Why the $39.79 Billion Forecast Matters to Peptide Outsourcing Decisions
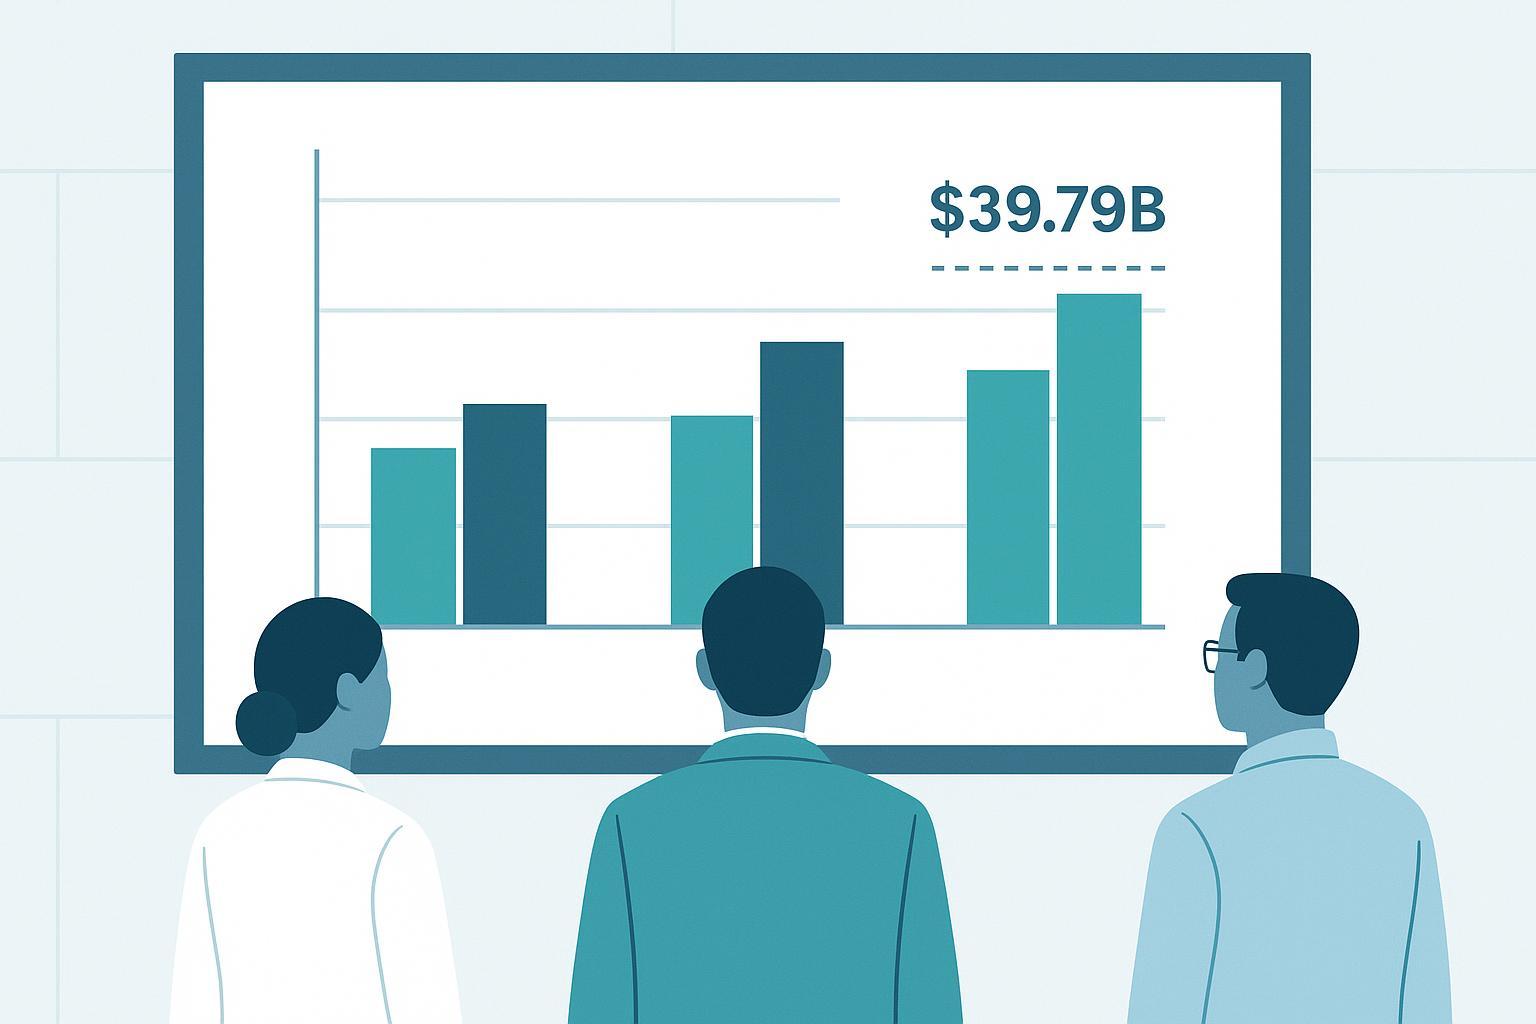
The drug discovery services market is often quoted as a single number, but the number changes with the definition. Research and Markets puts the 2030 figure at $39.79 billion from $23.21 billion in 2026, a 14.4% CAGR, counting revenue from pharmaceutical, DMPK, biological and medicinal chemistry services (Research and Markets, Drug Discovery Services Market Report 2026, accessed 2026). That chain is single-upstream: no other provider in this review reproduces it.
A narrower CRO-facing definition puts the 2030 market at USD 27.23 billion, up from USD 16.36 billion in 2025 at a 10.7% CAGR (MarketsandMarkets, Drug Discovery Services Market Report 2025–2030, base year 2024). The gap traces to the CRO-to-pharma definition behind the smaller number, which excludes service revenue booked outside that relationship. A third provider puts the 2035 market above USD 89 billion at a 13.95% CAGR (Precedence Research, Drug Discovery Services Market Size & Global Trends 2035, published 2026-08-19).
Key Takeaway: The $39.79 billion forecast rests on one upstream source. Forecasts of this type vary by provider because they measure different scopes.
If the market’s own size depends on how scope is defined, a peptide partner’s deliverable needs the same explicitness.
The Conventional View: Buy Synthesis, Assemble the Rest Yourself
The dominant advice is to buy each step from the vendor who does that step best. Source custom peptide synthesis from the lowest-cost qualified provider, then contract modification, labeling, purification and analytics separately, on the logic that specialists beat generalists on both price and depth.
That logic is not lazy. Solid-phase synthesis matured as a discrete, quotable service, and procurement systems were built around per-task quotes: a sequence, a purity grade, a price, a lead time. CDMO capability guides, vendor FAQ pages and market-report summaries all reinforce the model, and the market it serves is real. MarketsandMarkets values the peptide synthesis market itself at under a billion dollars today, USD 0.98 billion in 2026, reaching USD 1.54 billion by 2031 at a 9.5% CAGR. Grand View Research puts the 2024 base at USD 961.5 million growing to USD 1,840.6 million by 2033, and identifies the CDMO and CRO end-use segment as the fastest-growing part of peptide synthesis.
For a short, simple, well-behaved sequence, per-task sourcing genuinely wins. The critique that follows has to be specific about where it stops winning.
Where Fragmented Peptide Outsourcing Breaks Down
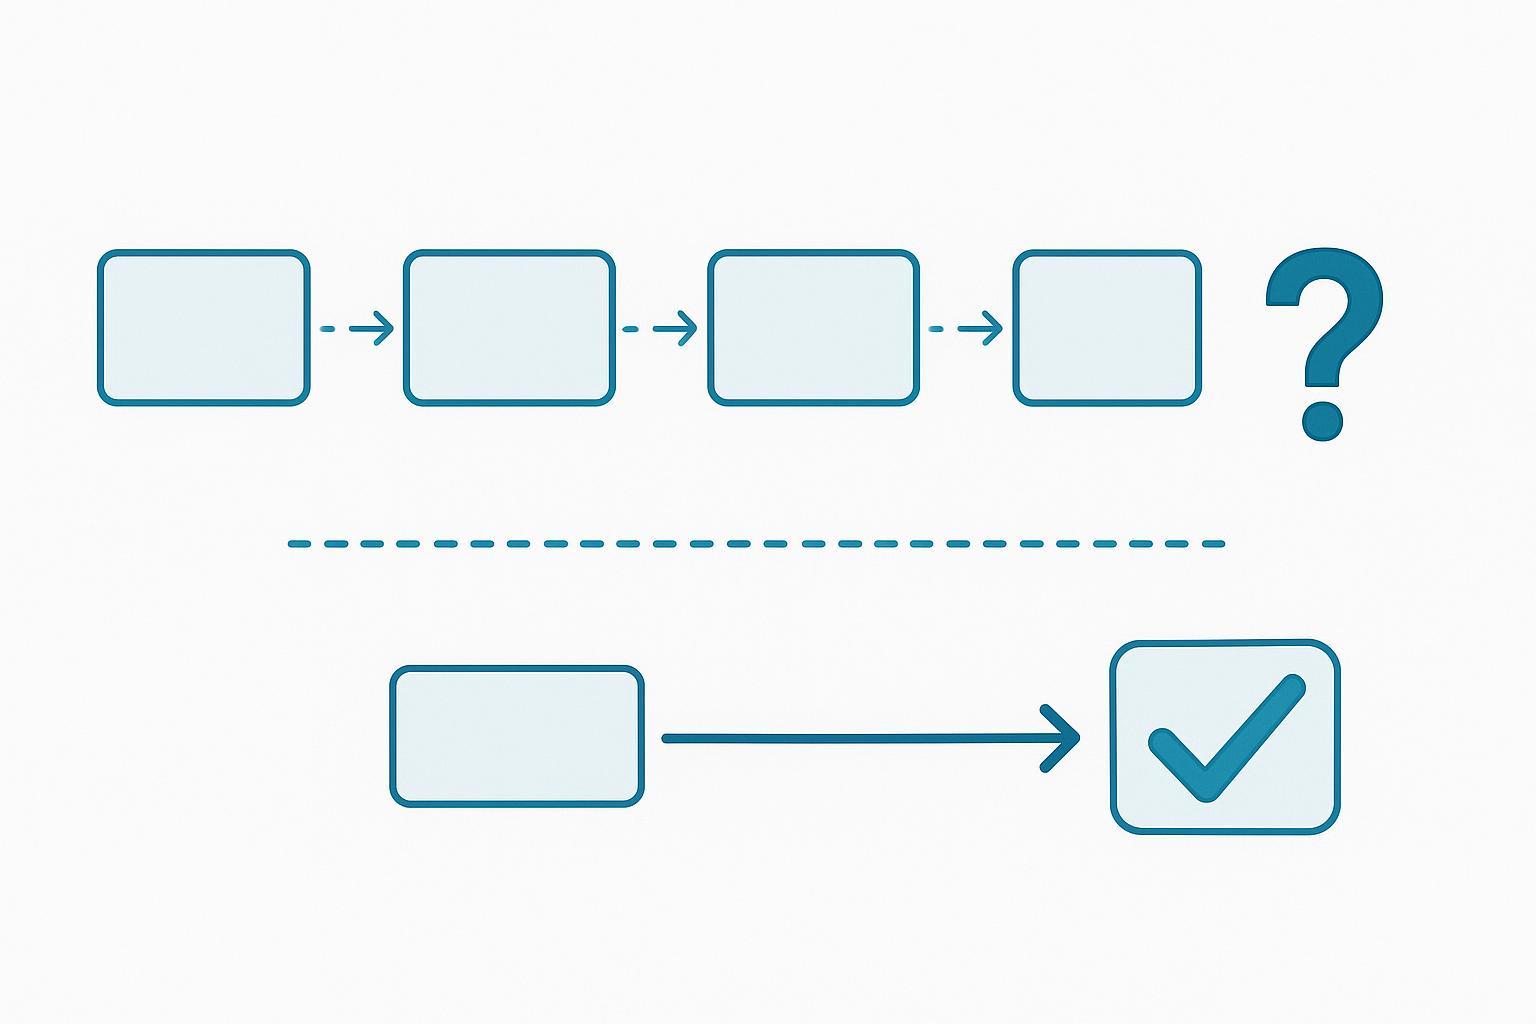
Fragmented peptide outsourcing does not remove integration work; it moves that work onto your team and breaks the traceability chain in the process.
The first break is the data chain. Each vendor’s certificate of analysis references its own batch ID, method version and acceptance criteria, so reconstructing one lot history across five suppliers becomes manual archaeology rather than a query.
Difficult sequences fail at the seams. Hydrophobic, aggregation-prone and multi-site modified sequences lose yield at every transfer, and above 95% purity at kilogram scale usually takes multi-step preparative HPLC, which means the recovery cost lands on whoever owns the final step. Recovery, not synthesis, is where most peptide material is lost: 50–80% crude yield, then 20–50% after purification.
Timelines compound rather than add. Adesis estimates that each added vendor handoff can add months of tech-transfer time, with multi-CDMO workflows stretching to 24–36 months. That is a vendor estimate, not a benchmark.
⚠️ Warning: Across vendors, delay and cost multiply. Six months per handoff does not stay six months when four handoffs queue behind each other.
The conventional model optimizes each task and suboptimizes the peptide outsourcing program.
What the Data Actually Shows: Integration Is the Variable That Moves
The forecasts everyone quotes measure spend, not outcomes. Grand View Research’s peptide therapeutics market report puts the peptide therapeutics market at USD 140.9 billion in 2025, rising to USD 294.6 billion by 2033 at an 8.7% CAGR, and that demand pressure lands upstream of synthesis, on teams that must move more sequences through modification, labeling, purification and testing than their internal capacity allows.
પેપ્ટાઇડ સંશ્લેષણ That is why vendor count is the wrong variable. For each program, ask whether the deliverable is a material or a decision. If it is a decision, require one accountable party across sequence design support, synthesis, modification, labeling, purification and analytical testing, which is what integrated peptide services actually means in practice.
Pro Tip: Ask whether the deliverable is a material or a decision. Materials can be sourced per task; decisions need one owner.
A batch-specific release package shows the difference concretely: the synthesis lot, the modification step and the analytical results carry one batch identifier and one method-version register, so a reviewer can trace a purity value back to the run that produced it.
The honest limit: no retrieved source quantifies integrated versus fragmented adoption as a percentage, so this argument rests on mechanism and documented artifacts, not on an adoption statistic.
What a Traceable Data Package Must Contain
A traceable data package is a lot-specific record that ties every result to a batch, a method version and a release decision. A certificate of analysis should tie every result to one lot, and if it does not, the number on the page is not evidence of anything.
Compiled vendor guidance, not a published standard, converges on the same minimum content. The table below consolidates it.
|
Required element |
What it must be tied to |
Failure mode when missing |
|---|---|---|
|
Batch/lot identifier |
The physical lot the sample came from |
Results cannot be traced to shipped material |
|
Identity and counterion or salt form |
The specified salt form for that lot |
TFA and acetate lots look interchangeable on paper |
|
Analytical method identifiers and versions |
The exact method revision used |
A method change silently invalidates comparability |
|
Acceptance criteria |
Stated before results, not after |
Specifications get written to fit the result |
|
Purity result with integrated chromatogram and peak-area table |
The integration, not a summary percentage |
A single number hides co-eluting impurities |
|
Raw RP-HPLC chromatograms |
The run that produced the purity figure |
No way to re-integrate or audit the baseline |
|
LC-MS spectrum with observed versus theoretical mass and charge states |
The same lot and method version |
Identity is asserted rather than demonstrated |
|
Impurity profile, specified, unspecified and total |
The reporting threshold applied |
Unspecified impurities go uncounted |
|
Peptide mapping, where used |
The digest and column conditions |
Sequence confirmation cannot be reproduced |
|
Counterion confirmation |
The lot’s declared salt form |
Stoichiometry and assay drift |
|
Endotoxin by LAL, sterility or bioburden, where required |
The release specification for the intended use |
A research-grade lot is treated as clinical-grade |
|
Deviation, OOS and OOT records with QA disposition |
The batch record and the disposition decision |
Failures disappear between testing and release |
|
સેવાઓ Testing lab and approver |
Named laboratory and named approver |
Accountability is unattributable |
|
Final released or withheld decision |
The lot, dated |
Material ships without a documented release |
Two named references frame the analytical side. ICH Q2(R2), effective June 2024, requires validation of analytical procedures for their intended use, so a method identifier without a validation status is an incomplete record. USP <71> and USP <85> are the relevant compendial references for sterility and bacterial endotoxins; their clause wording is not reproduced here. ALCOA data-integrity expectations apply to the raw data, the integrations and the audit trail, not only to the reported summary.
Purity specifications move with program stage, and these bands are vendor-sourced rather than harmonized. Research-grade material is commonly 70%, with 80, 90, 95 and 98% options selected by assay use. Preclinical work generally runs 90 to 98%, with at least 95% recommended for early screening and IND-enabling studies. Clinical material is generally at least 98%, with single impurities held below 0.5%.
The practical test is simple: ask for the package before you ask for the price. A vendor who cannot produce the integration behind the purity figure has told you what the rest of the record will look like.
Red Flags and Must-Haves in Peptide CRO Selection Criteria
Red flags cluster around documentation that cannot be traced. A certificate of analysis that reports a purity percentage with no integrated chromatogram or peak-area table is a red flag. So is a result that is not tied to a method version, an acceptance criterion that appears only after the results are in, and a batch that underperforms without a deviation or out-of-specification record. A single accountable contact who cannot produce raw data on request belongs on the same list, as does a partner who quotes custom peptide synthesis alone and leaves modification, labeling, purification and analytics as your coordination problem.
Must-haves are the mirror image, and each one is checkable. Ask for one batch identifier spanning every stage, a method-version register, and orthogonal characterization at minimum by RP-HPLC plus LC-MS rather than a single method. Ask for the documented counterion and salt form, the named approver and testing lab, and a written release decision. Then attach a scenario: when a 40-mer hydrophobic sequence needs on-resin aggregation control and the purification step sits with a different vendor, what happens to the batch record? If nobody can answer, the traceable data package is a promise rather than a process.
One caution on specifications. Vendor pages commonly publish a purity ladder, with research-grade material around 70% and 80/90/95/98% options, preclinical work roughly 90-98% and often ≥95% recommended for early screening and IND-enabling studies, and clinical material generally ≥98% with single impurities below 0.5%. Those figures come from vendor specification pages, not from a regulator, so treat them as a starting point: purity grade is a specification, not a marketing label, and your program’s regulatory pathway should set the acceptance criteria you actually write into the contract.
How to Apply the Criteria to a Decision
Pull the last three CoAs from your current partner and check one thing: is every result tied to a batch ID and a method version? If not, you have found your starting point.
-
Write your acceptance criteria and required data-package contents before contacting any vendor. Under a day, and it is the highest-leverage step in the sequence.
-
Map which stages your current model outsources separately, and mark every Shop handoff where the batch identifier changes. One to two weeks.
-
Send the checklist to two or three candidate partners and score responses on artifact completeness, not on price or claimed capability. કૃત્રિમ પેપ્ટાઇડ્સ When you compare integrated peptide services, ask which batch ID follows the material across synthesis, modification and analytics. Two to three weeks.
-
Require a written release decision and a named approver in the contract. Longer-term, but it is what converts a promise into a process.
-
Re-audit the first delivered lot against the checklist from step 1.
The deliverable you are scoring looks like this: a batch-specific release package where the synthesis lot, the modification step and the analytical results share one batch identifier and one method-version register, with raw chromatograms supplied alongside the summary result. That is the artifact, not a claim about it.
Track two numbers as you go: hours your team spends reconstructing lot history per program, and batches requiring re-analysis. Those are the costs fragmentation hides. Expect the first fully traceable package on your next program cycle, not the current one.
Next step: Request a feasibility assessment and ask MOL Changes to show you what a batch-specific release package looks like for a sequence like yours.
Caveats: Where the Conventional Model Still Wins
The strongest objection to this argument is that integration is not always worth paying for. For a single, well-characterized, unmodified sequence at small scale, per-task sourcing can be cheaper and faster, and consolidation adds coordination overhead that a one-off order does not need.
Context matters in a second way. A team with an established internal analytical function may already own the traceability chain, so it gains less from buying that chain as a service.
The weakest part of the case is evidential. No retrieved source quantifies the cost of fragmentation or the adoption rate of integrated models, so the argument rests on mechanism, documented artifacts and vendor estimates rather than a controlled comparison.
The bounded position: integration earns its premium when the program’s deliverable is a decision and the sequence is difficult, modified or labeled. When the deliverable is simply a material, it is optional.
But doesn’t per-task sourcing give better depth and price?
Often, yes, for a single unmodified sequence. If your program needs one 20-mer synthesized to 95% purity and nothing else, a specialist shop that does only that will usually beat an integrated provider on both unit price and depth of expertise, because you are buying exactly one capability and paying for exactly one capability.
The calculus flips once the sequence needs modification, labeling or orthogonal characterization. At that point the work splits across vendors, and the buyer absorbs the integration labor: reconciling batch IDs, chasing method versions, and re-running characterization because one supplier’s certificate does not answer the next supplier’s question. That labor rarely appears in the per-task quote, which is why the cheaper line items can still produce the more expensive program. વિશે
What if we have already built a multi-vendor peptide supply chain?
You do not have to replace every vendor to fix this. The first move is a documentation change, not a re-sourcing decision: impose one batch identifier across all existing suppliers, so that a lot from one vendor and a lot from another can be referred to by the same internal name.
Then request, retroactively, the method versions and raw data behind the current lot. Most suppliers will release this when asked directly, and what they cannot produce tells you where the risk sits. Apply the full checklist at the next program cycle rather than mid-program, and consolidate vendor by vendor as contracts come up for renewal. That way the traceable data package grows around work already underway instead of restarting it.
How do you respond to the market reports that show outsourcing growing steadily?
Steady growth in peptide outsourcing is real, and it is consistent with this argument rather than against it, because those reports measure spend, not traceability. MarketsandMarkets puts the peptide synthesis market itself under a billion dollars today, at USD 0.98 billion in 2026 rising to USD 1.54 billion by 2031 on a 9.5% CAGR from a 2025 base year. Grand View Research reaches a similar order of magnitude from a different base, USD 961.5 million in 2024 to USD 1,840.6 million by 2033 at 7.71%, and finds the CDMO and CRO end-use segment is the fastest-growing part of peptide synthesis. Figures vary by provider and by what each report counts: reagents, consumables, and custom peptide synthesis across liquid-phase and solid-phase routes.
The two estimates do not agree, and that disagreement is the point. MarketsandMarkets frames drug discovery services through a narrower CRO-facing definition, early-phase work performed by contract research organizations, while services-revenue estimates count a broader set of activities. A growing number therefore tells you demand is rising. It does not tell you what a given partner will hand back with your lot. પેપ્ટાઇડ ઉત્પાદન
Pro Tip: Ask a prospective partner which market definition they benchmark against. A CRO-scope number and a services-revenue number are not comparable, and neither one tells you what your lot record will contain.
Next Steps for Peptide Teams Evaluating Integrated Services
The forecast is a reason to define the deliverable, not a reason to buy more tasks. Start your peptide outsourcing review by writing the acceptance criteria and the required data-package contents before you contact any vendor: a checklist written after the quote is a negotiation, not a specification.
-
Draft your release-package requirements today (under an hour). Ask for a lot-specific release package that ties every result to one batch, and name the analytical procedures you expect to see validated under ICH Q2(R2), effective June 2024.
-
Map your current program against the criteria in this article and mark every gap.
-
Request one accountable party across sequence design support, synthesis, modification, labeling, purification and analytical testing, so integrated peptide services replace handoffs between vendors.
-
Ask each candidate for a sample traceable data package, including raw chromatograms and method versions alongside the summary result. MOL Changes is one example of a partner that supplies this artifact; the deliverable, not the vendor name, is what you score.
Measure progress by whether the next program cycle produces a package you can audit without emailing the lab. Expect the first fully traceable release on that cycle, not this one.
Key Takeaway: Write the acceptance criteria and the required data-package contents before you contact any vendor, a checklist written after the quote is a negotiation, not a specification.
Pressure-test your next peptide program before you commit. Send us your sequence and target specification; we will return a feasibility assessment and a sample analytical package so you can score the deliverable against your own checklist. Request a feasibility assessment.
Disclosure: MOL Changes has a commercial interest in peptide quality standards. This article covers research-use materials and program decisions only; consult a qualified professional before making medical or regulatory decisions.