What did the FDA actually approve or publish in October 2026?
The FDA’s own monthly drug index for October 2026 records three oncology-related approvals and two newly added guidance documents in the 1 に 13 October window. The approvals were pirtobrutinib on 2 10月, tucatinib on 7 October and atezolizumab on 8 10月; the guidances were the ICH E2B(R3) electronic case-safety-report implementation guide on 1 October and the postapproval manufacturing changes guidance on 9 10月 (FDA What’s New Related to Drugs, page current 2026-10-13).
The tucatinib approval notification is the clearest example of how narrow these indications have become: tucatinib with trastuzumab and pertuzumab for maintenance treatment of adults with unresectable locally advanced or metastatic HER2-positive breast cancer, limited to patients with no disease progression after 4 に 8 induction cycles of trastuzumab, pertuzumab and a taxane (FDA tucatinib approval notification, 7 10月 2026).
The one guidance with transferable weight is the guidance FDA finalized on 9 10月 2026 on postapproval manufacturing changes to biosimilar and interchangeable products, a Level 1 final guidance whose Federal Register notice is dated 13 10月 2026 (FDA guidance page, issued October 2026).
スケール用, FDA counted 46 novel drugs approved in 2025 under its own definition of never before approved or marketed in the US (FDA Novel Drug Approvals for 2025, page current 2026-07-08). That figure excludes new indications, which the agency reports separately, so it cannot be read as a peptide count.
Which of those actions change what a peptide developer must measure or document?
One item changes measurement and documentation practice for peptide developers, and it does so by transferable logic rather than by direct scope: the guidance FDA finalized on 9 10月 2026 (FDA guidance page, issued October 2026).
The reason is that comparability has a defined meaning. ICH Q5E’s definition of comparable requires pre- and post-change product to be “highly similar” with no adverse impact on safety or efficacy including immunogenicity, and it states plainly that comparability does not require identity of quality attributes; where analytical procedures cannot discern relevant differences, additional testing or nonclinical and clinical studies should be considered (I Q5E, ステップ 4).
ペプチド 2 工場 Peptides complicate that because they sit between two quality regimes. The EMA’s synthetic-peptide guideline states that synthetic peptides are fully or partially excluded from the scope of ICH Q3A/Q3B, Q6A/Q6B and M7, and that they sit at the interface of small molecules and proteins (EMA合成ペプチドガイドライン, 2022).
I Q3A(R2) excludes peptides from its direct scope, so its small-molecule thresholds are not default peptide release limits. Establish the appropriate peptide impurity framework and product-specific justification.
What must a peptide-specific analytical package contain, and how do the records connect?
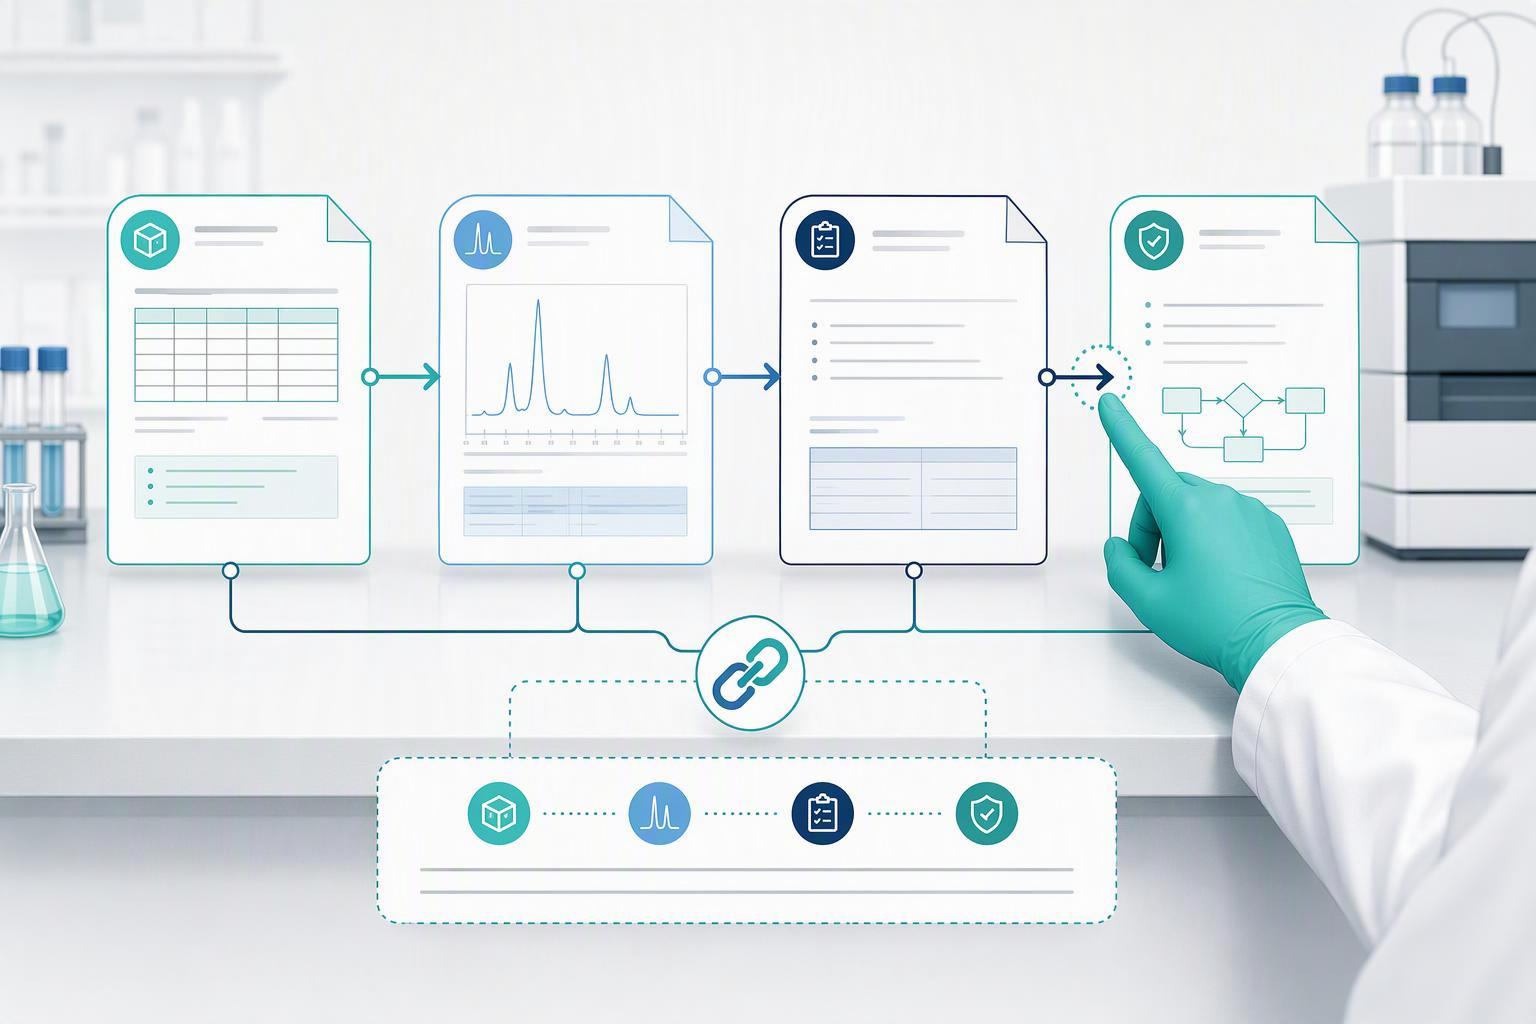
A peptide analytical package is a specification plus the validated methods and records that make each number defensible, not a purity figure on its own. The Q6B specification framework defines a specification as a list of tests, 分析手順と合格基準への言及, requires identity tests to be highly specific and ペプチドサプライヤー サプライヤー based on unique aspects of molecular structure, permits more than one identity test, and states that absolute purity is difficult to determine and method-dependent, so purity is estimated by a combination of methods (I Q6B, ステップ 4 1999).
The methods behind those tests have to be validated against the Q2(R2) validation characteristics: specificity and selectivity including orthogonal procedure comparison, range including the lower range limit and LOD/LOQ, accuracy and precision including orthogonal comparison and spiking, と堅牢性, with linearity handled within range using a minimum of five concentrations (I Q2(R2), ステップ 4 バージョン 2023-11-30).
The records then have to survive inspection. 21 CFRパート 11 treats electronic records and signatures meeting its criteria as equivalent to paper, applies to records created, modified, 維持された, archived, retrieved or transmitted under agency records requirements and to records submitted to FDA, and requires computer systems, controls and documentation to be readily available for, and subject to, FDA inspection (21 CFRパート 11, 最新版).
Sterility and bacterial endotoxin tests answer different questions. Include them when justified by dosage form, ルート, intended use and applicable requirements, using current methods and demonstrated suitability rather than treating the pair as universal peptide release tests.
When synthesis site, 精製, formulation or scale changes, what comparability evidence is expected?
Best Peptide Synthesizer A comparability package after a manufacturing change is expected to show the pre- and post-change product are highly similar, with side-by-side data rather than a single post-change lot. ICH Q5E’s definition of comparable sets that bar and states that where analytical procedures cannot discern relevant differences, additional testing or nonclinical and clinical studies should be considered (I Q5E, ステップ 4). ペプチドサプライヤー工場
質問1A(R2) stability designs depend on product, storage and submission context. For the general case, long-term conditions may be 25 °C/60% RH or 30 °C/65% RH; intermediate testing is conditional, not a universal additional six-month requirement. Research reagents need a justified use-specific stability plan rather than automatic adoption of a registration data package. 質問1A(R2).
Any cited four-month BsUFA review goal must be read in its specified supplement category and performance-goal context. Beta Endorphin Synthesis It is not a guaranteed decision date or a general duration for preparing or reviewing a synthetic-peptide package.
How do narrow indications and combination regimens constrain peptide program design and labeling?
Narrow indications and combination regimens push the analytical burden earlier, because the label has to describe a population defined by prior treatment and biomarker status. The tucatinib approval notification ties the indication to patients with no disease progression after 4 に 8 induction cycles, which makes the prior-therapy history part of the labeled population rather than a clinical detail (FDA tucatinib approval notification, 7 10月 2026).
That pattern is not confined to one product. A retrospective review of the full FDA oncology approval set from 2011 に 2023, カバーリング 468 approvals and 238 製品, それを見つけた 40% of regular and 48% of accelerated cancer therapeutic approvals were for biomarker-defined populations (Biomarkers for Precision Patient Selection in Cancer Therapy Approvals in the US, 2024). Those shares are oncology-specific and should not be read as all-drug figures.
ペプチドプログラムの場合, the practical consequence is that the specification has to support the labeled population from the start, which is why the identity and impurity work in the analytical package is not separable from the labeling decision. Branched Peptide Synthesis
Which October headlines do not touch peptide programs at all?
The July 2026 compounding votes are background, not October news. The July 2026 PCAC meeting recommended adding BPC-157, KPV, TB-500, c-WORDS, Semax and Epitalon to the 503A Bulk Drug Substances List and rejected emideltide/DSIP; the votes are advisory and nonbinding and, as of October 2026, none of the six had been formally added (FDA PCAC meeting page, meeting 2026-07-23/24).
よくある質問
Did the FDA issue any peptide-specific guidance in October 2026?
The selected FDA records reviewed through 15 October do not establish an exhaustive absence of peptide-related actions for the month. This article discusses the named updates and their scope, including the July withdrawal of the 2021 合成ペプチドガイダンス.
Does the October 2026 biosimilar manufacturing guidance apply to my peptide?
Only if your product is licensed as a biosimilar or interchangeable biological product. A synthetic peptide reviewed as a drug under the FD&C Act falls outside that scope. The useful part is the reasoning, not the applicability: the guidance’s risk-based approach to post-change evidence tracks ICH Q5E’s definition of comparable, which asks for pre- and post-change material to be highly similar with no adverse effect on safety or efficacy, immunogenicity included. Treat Q5E as the transferable standard.
Do the July 2026 compounding votes mean these peptides are now legal to compound?
いいえ. The July 2026 PCAC votes are advisory and nonbinding, and as of October 2026 none of the six substances had been formally added to the 503A Bulk Drug Substances List. Keep research-use framing distinct from any human-use or approval claim, and confirm supplier language avoids dosing, therapeutic, or diagnostic claims.
結論
A useful package links specifications to appropriate methods, actual lot data and review decisions. Determine validation characteristics, stability design and electronic-record controls from the product, stage and jurisdiction. No single Q6B or general stability template is a complete specification for every synthetic peptide.
The next step is a gap check against that list before your next change, それ以降ではない. If you want a second read on an analytical package or a comparability plan, that is a documentation conversation rather than a filing one, and it is the right time to have it.