Why one atom rewires biology
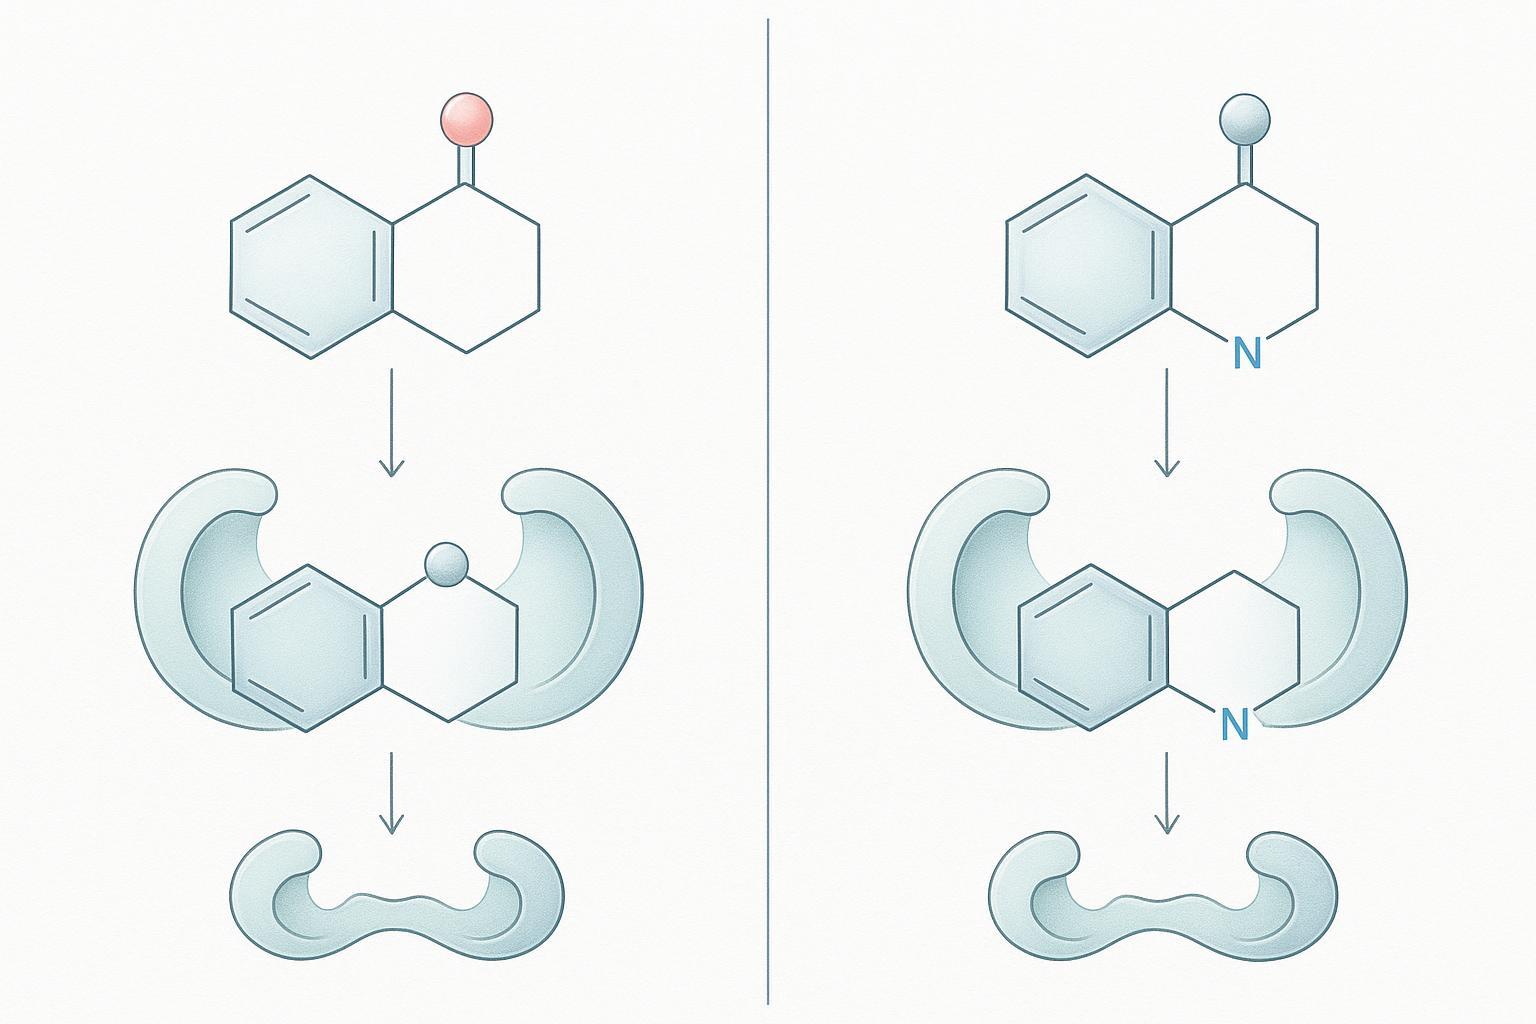
That redirection is not a subtle shift in degree. In 2026 skeletal-editing work, an oxygen-to-nitrogen edit converted limonin’s furan ring into a pyrrole, producing the analog 15N-LIMO. In a thermal binding assay plus cell studies, limonin preferentially bound kinases while the edited analog bound a different protein set including nicotinamide nucleotide transhydrogenase (NNT). Both compounds were antifibrotic, but through different mechanisms: limonin inhibited signaling pathways, while 15N-LIMO acted on mitochondrial redox and reduced ROS. 15N-LIMO was also less cytotoxic. The project, led by Yoonsu Park at KAIST with Ho Jeong Kwon at Yonsei, is described as a dramatic rewiring of target engagement (C&EN, 2026-09-15).
Two boundaries matter here. Accessible reporting gives no single potency value for the shift, so none is claimed. And limonin is a citrus natural-product-like small molecule, not a peptide or depsipeptide, which makes the transfer to peptide-like scaffolds an argument by analogy rather than a peptide result (C&EN, 2026-09-15). A related 2026 advance, diastereoselective ring expansion of tetrahydrofurans to pyrrolidines, shows the same oxygen-to-nitrogen logic applied at ring level (Nature Communications item, 2026-02-02).
The implication for modified peptide characterization is direct: if one atom can change what a molecule binds, a purity percentage cannot stand in for identity.
The conventional view: purity as the gate
Sintéis Peptide If one atom can change what a molecule binds, the question becomes what the release gate actually measures. The mainstream gate for a peptide-like molecule is a set of impurity thresholds, and both major regulators still frame it that way. The FDA’s July 2026 revised draft guidances for generic peptide products keep the familiar reporting, identification and qualification tiers at 0.10%, 0.50% and 1.0%, with the draft lowering the reporting threshold to 0.05% for products dosed above 10 mg per day. The EMA synthetic peptide guideline that took effect in June 2026 sets reporting above 0.1%, identification above 0.5% and qualification above 1.0%, and controls those limits against the Ph. Eur. general monograph “Substances for Pharmaceutical Use” rather than ICH Q3A, which does not apply to synthetic peptides.
Peptides sintéiseacha src=”https://molchanges.com/wp-content/uploads/2026/09/pub_20260925_050922_934_21bf4c59c3ab42f2ab0ac9a6e982f6ba.png”>
|
Framework |
Seirbhísí Reporting |
Identification |
Qualification |
Basis |
|---|---|---|---|---|
|
FDA (July 2026 draft) |
0.10% (0.05% above 10 mg/day) |
0.50% |
1.0% |
Product-specific guidance |
|
EMA (effective 2026-06-01) |
>0.1% |
>0.5% |
>1.0% |
Ph. Eur. “Substances for Pharmaceutical Use” |
That framework became the default because it is auditable, comparable across lots, and far cheaper than orthogonal work. Note that the FDA landing page names impurity thresholds without printing the numbers, so the FDA figures above are secondary-sourced.
Why HPLC purity passes while identity is wrong
The reason those thresholds cannot carry the identity question is that an HPLC purity number reports chromatographic homogeneity, not chemical identity. A single peak area tells you one thing only: the material eluted as one band under those conditions. It does not tell you what that band is.
Three failure modes exploit that gap. First, structurally different peptides can share near-identical retention and appear as one peak, so sequence isomers inflate apparent purity without ever being separated; as a standard peptide-analysis reference puts it, a peak area is not an identity (HPLC Analysis and Purification of Peptides, undated, retrieved 2026-06-14). Second, coupling-stage racemization via a planar oxazolone intermediate lets the incoming amine attack either face of the activated residue, inverting D/L stereochemistry at a single position; His and Cys are the classic sensitive residues, and because epimers are isobaric with the target, mass alone cannot resolve them (EMA guideline on the development and manufacture of synthetic peptides, effective 2026-06-01). Third, truncation and deletion sequences from incomplete coupling co-elute or surface only as shoulders, while oxidation at Met, Trp or Cys and deamidation at Asn or Gln shift retention so subtly that they hide inside the main peak (HPLC Analysis and Purification of Peptides, undated, retrieved 2026-06-14).
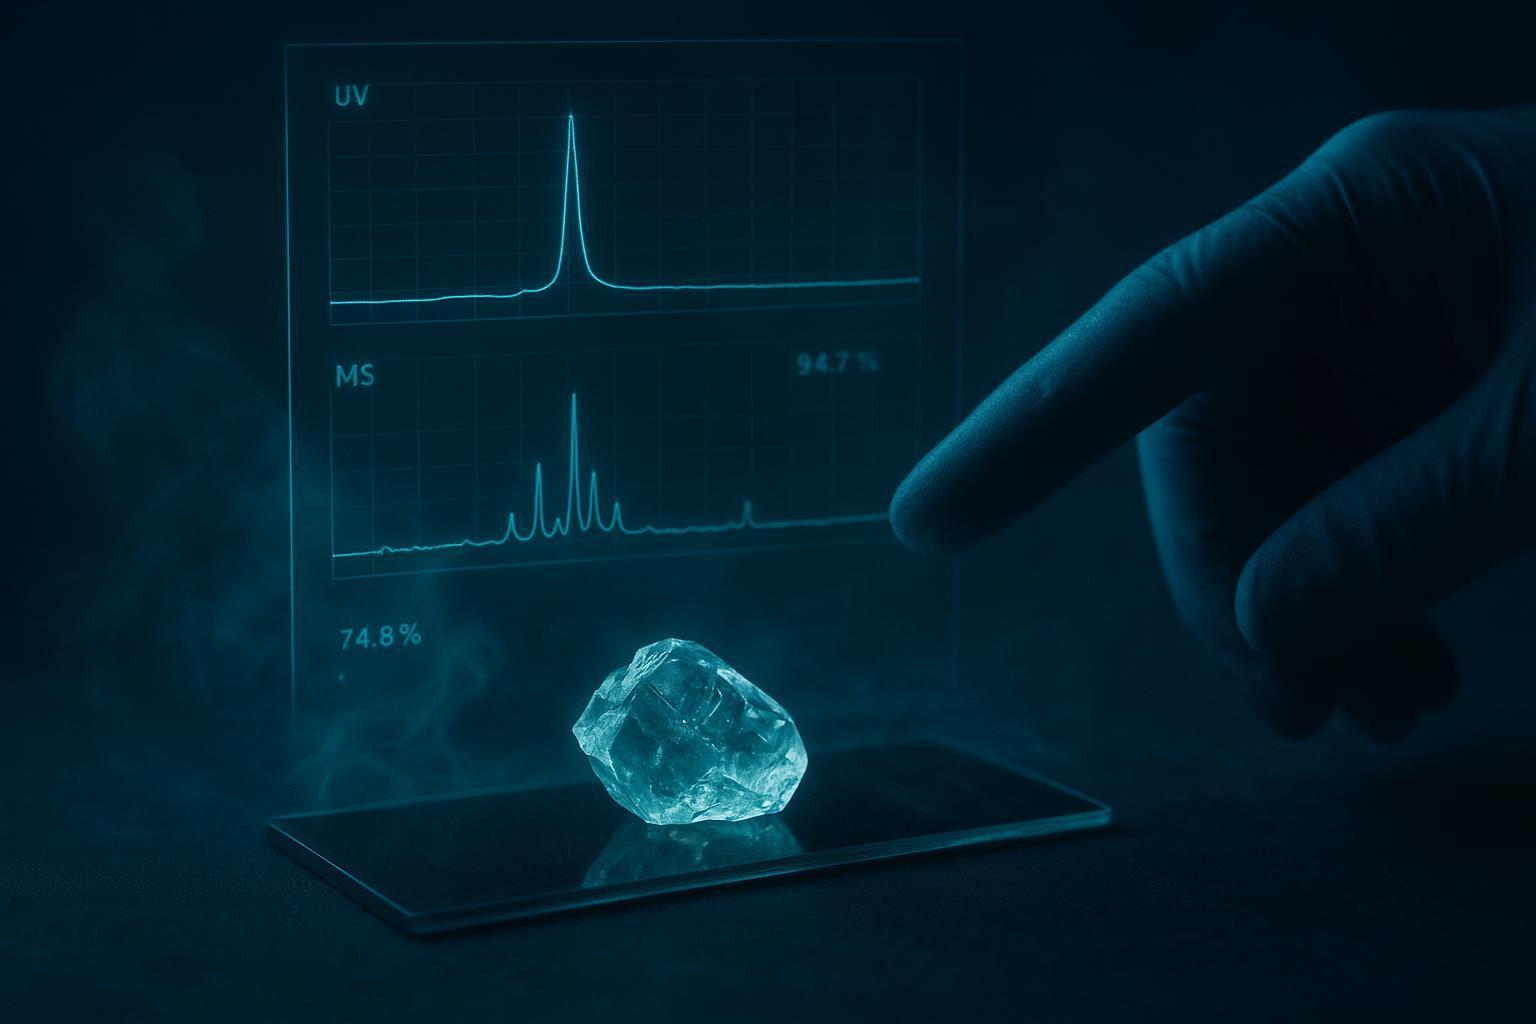
The magnitude is not hypothetical. One vendor’s own comparison of UV and MS purity on the same Eledoisin lot reported 94.7% by optical detection against 74.8% by mass spectrometry, because co-eluting peptides appear only in the MS trace (Waters application note, 2018-08-01). Waters sells LC-HRMS, so read that number as a vendor making its own case, not as a neutral benchmark.
⚠️ Rabhadh: A passing purity specification is not evidence of peptide structural confirmation. It is evidence that your detector saw one peak.
All three modes share one property: they are invisible to a detector that only counts peak area.
What the data actually shows about modified peptide characterization
Because those failure modes are invisible to peak-area detection, the reframe is simple: the useful question is not “how pure is it” but “which method answers which question.” Once you ask it that way, the answer set is orthogonal by construction, and no single release test can cover it.
A 2025 study on four cyclic nonapeptides found that RP-HPLC and HILIC resolve different impurity populations from the same samples, so each separation sees a fraction the other misses (Journal of Chromatography A, 2025). That is the practical case for pairing methods rather than ranking them.
Detection limits are real but narrow. One LC-HRMS workflow reported an API-Proline impurity at 0.08% relative abundance with 2.4% RSD, a single-peptide, instrument-specific result rather than a general floor (Waters application note, 2018).
The mechanism explains why the O→N case resists a purity-only read. An amide nitrogen supplies a backbone N–H hydrogen-bond donor and a strong carbonyl acceptor; swap it for oxygen and the ester cannot donate that hydrogen bond and its acceptor carbonyl is weaker (Peptide-Bond Isosteres review, 2011). A specific intra- or intermolecular hydrogen bond is lost, the backbone becomes more flexible with a lower rotational barrier, and the cis/trans ensemble shifts. The target then sees a different population of conformers, not a different mass.
Direction matters too. Amide→ester removes a donor and adds flexibility; ester→amide restores the N–H donor and a stronger acceptor, tightening conformational bias (Amide Bond Bioisosteres review, 2020).
So a mass-confirmed, high-purity sample can still present the wrong conformer population. Structural confirmation and peptide bioactivity interpretation are separate questions from purity, and the EMA expects orthogonal methods for identity, including higher-order structure where relevant (EMA guideline explainer, 2026). ICH Q2(R2) supports this by letting a second well-characterized orthogonal procedure carry the specificity argument when impurity standards are unavailable (ICH Q2(R2), 2023).
Key Takeaway: Purity and mass confirmation do not establish which conformer population your sample presents. That is a structural question, answered by orthogonal methods, not by a higher purity number.
The better approach: an integrated characterization package
If conformer population is a structural question, the package has to be built to answer it. Treat synthesis route, impurity map, structure, and bioactivity as one linked package rather than four checkpoints. The core principle: every claim must be anchored to a named standard or documented practice, and every method’s limits must be stated before its result is used.
Five rules make that work.
-
Define the question before the method. Íonachta, identity, conformation, and function are different questions. Choosing the instrument first guarantees you answer the wrong one.
-
Require orthogonality, not redundancy. Two methods that separate by the same mechanism confirm each other’s blind spots. If reversed-phase HPLC and a second reversed-phase gradient both miss a co-eluting variant, running both adds cost, not confidence.
-
Treat the impurity map as a structural hypothesis. The EMA’s explicit impurity taxonomy names truncated, deletion, insertion, racemization-derived stereoisomer, oxidation, and deamidation products as distinct categories, each with its own analytical consequence. A plain-language walkthrough of what the guideline asks for is useful for translating that taxonomy into a testing plan.
-
Connect structure to function with a bioassay that distinguishes mechanism. A potency readout that cannot separate receptor affinity from downstream signaling will not tell you whether a single-atom change altered binding or altered signaling.
-
State limitations up front. Chiral methods consume more material and time; ion mobility resolution depends on the variant’s collision cross-section. Naming those limits before the result is used is what makes the result defensible.
For the package as a whole, ICH Q2(R2) validation expectations set the bar for demonstrating that each method performs as claimed, and the FDA’s Shop July 2026 revised draft guidances for generic peptide products show regulators moving toward the same structural-identity expectations.
Chun Leid: Write the impurity map as a set of structural hypotheses, not a list of peak areas. Each named species implies a synthesis or handling step you can then test directly.
A co-eluting isobaric variant sitting under the main peak is the case that exposes a purity-only workflow. MS detects the mass difference first; chiral analysis then resolves which stereoisomer is present. MOL Changes supports that kind of sequence, and the same logic can be used to decide which orthogonal method answers the question the first one raised.
|
Project scenario |
Recommended characterization scope |
|---|---|
|
Modified or conjugated peptide |
Impurity map against the EMA taxonomy, MS/MS connectivity, bioassay with mechanism resolution |
|
Cyclic or disulfide-rich scaffold |
Disulfide pairing assignment, conformational check, orthogonal separation |
|
Lot-to-lot comparability |
Same method set across lots, with defined acceptance criteria per attribute |
|
Potency shift with unchanged purity |
Chiral or isobaric resolution first, then structural confirmation before re-testing potency |
How to apply this
Putting the package together starts with writing down the question your data must answer and the decision it feeds. If the release decision is “does this batch match the intended molecule,” a purity percentage is not an answer. This takes an hour and usually exposes the gap immediately.
Then audit the current package for redundancy. Two methods that separate on the same principle, reversed-phase and another reversed-phase gradient, answer one question twice. The EMA guideline on synthetic peptides is useful here: its impurity categories let you rank failure modes by risk for your specific scaffold, so you add the orthogonal method that covers the highest-risk one rather than the easiest one to run. Expect this step to cost instrument time and material. Maidir
Build the impurity map into a structural hypothesis, then test it against the bioassay. That feedback loop is where peptide bioactivity interpretation stops being a number and becomes a mechanism. Validate each method for the role it plays under ICH Q2(R2), and document its blind spots next to the result.
Track one thing: whether identity, stereochemistry, conformation and bioactivity each have at least one method behind them.
Caveats
The strongest limitation is that the O→N evidence comes from a natural-product-like small molecule, not a peptide, so the mechanism transfer is analogical rather than demonstrated, and the strength of that analogy varies by scaffold. The natural-product scope of the original finding is the reason to treat it as a warning about single-atom change peptide biology, not as a peptide-specific result.
For a short, unmodified peptide made by a mature route, the conventional purity gate may be sufficient, and the added orthogonal work may not change a release decision. The mechanistic basis for the conformational shift and the directionality of the amide–ester swap support the argument, but no peptide dataset in the accessible literature confirms that a shifted ensemble reliably produces a different biological outcome. That is the weakest link in this position.
The claim here is narrow: purity is necessary but not sufficient. It is not an argument for reducing purity testing.
But doesn’t the conventional purity gate work for well-behaved peptides?
Yes, for a short unmodified peptide on a low-risk scaffold, a purity-only release specification is often defensible, and nothing here argues for replacing it wholesale. The objection fails only when a passing purity figure is read as a statement about identity. A peak area measures how much material elutes as one band under one set of conditions; it does not establish connectivity, and a peak area is not an identity (PMC). That distinction is exactly what the guideline’s orthogonal identity expectation formalizes: purity and structural confirmation are separate questions, answered by separate methods (ChemVerify).
So the practical rule is proportional, not absolute. Keep the reduced package where the scaffold and the decision stakes justify it, but document the blind spots in the specification itself, so nobody downstream treats a clean chromatogram as proof of peptide structural confirmation.
What if we have already validated a purity-only release specification?
Keep the validated specification as your release gate. You do not have to reopen a filed method or renegotiate a specification to strengthen characterization. Add orthogonal work as a characterization-layer activity that sits alongside release testing rather than replacing it. Táirgeadh Peptide
The EMA’s guideline on synthetic peptides is explicit that co-eluting impurities must not be left as unresolved single peaks where they can be separated or otherwise justified. That single sentence is your prioritization rule: start with the failure mode most likely for the scaffold in front of you, not with a wholesale method rewrite. A scaffold prone to deamidation gets an orthogonal separation; one prone to sequence variants gets fragmentation-based confirmation.
Regulators are moving the same direction. Analysis of FDA expectations and trade reporting concludes that manufacturers relying solely on HPLC-UV will need to upgrade. Treat that as a documentation roadmap, not a verdict on work already done.
How do you reconcile this with the FDA and EMA thresholds?
The thresholds define what you must report, identify, and qualify. They do not define what you must structurally confirm. That gap is where a purity-only release specification quietly stops short of the guidance it claims to satisfy.
The FDA’s July 2026 revised draft guidances for generic peptide products keep the impurity reporting, identification, and qualification framework that has governed peptide submissions, and the FDA landing page names impurity thresholds without printing the numbers, so any specific figure should be checked against the current guidance text rather than a summary. The EMA synthetic peptide guideline that took effect in June 2026 sets its expectations on the same foundation, and the guideline itself ties impurity control to the active substance’s defined structure. Neither document asks a chromatographic purity value to stand in for structural identity. They assume you already know what the molecule is. Peptide impurity profiling answers the reporting question; it does not answer the identity question, and the two are not interchangeable in a filing.
Conclusion
Purity is necessary but not sufficient, because a single oxygen-to-nitrogen edit can change what a molecule binds: a single oxygen-to-nitrogen edit rewired a natural product’s protein targets entirely, which is why modified peptide characterization has to answer identity, connectivity, conformation, and bioactivity questions rather than report one number. The broader shift is a change in how packages get assembled. Instead of running a default method list and hoping the gaps do not matter, teams should build the package from the questions the molecule raises, and state openly what each method cannot see. A field where every reported purity figure travels with its method and its blind spots would waste less material, fewer retests, and fewer late surprises.
Next steps. If you are specifying a characterization package for a modified or peptide-like molecule and want the method set mapped to your actual structural risks, talk to our analytical team about an integrated characterization package covering impurity profiling, structural confirmation, and bioactivity interpretation.
Disclosure: MOL Changes provides analytical and synthesis services for modified peptides and related molecules, so we have a commercial interest in how these packages are specified.