Why the Burden of Proof Moved to Peptide Supplier Qualification
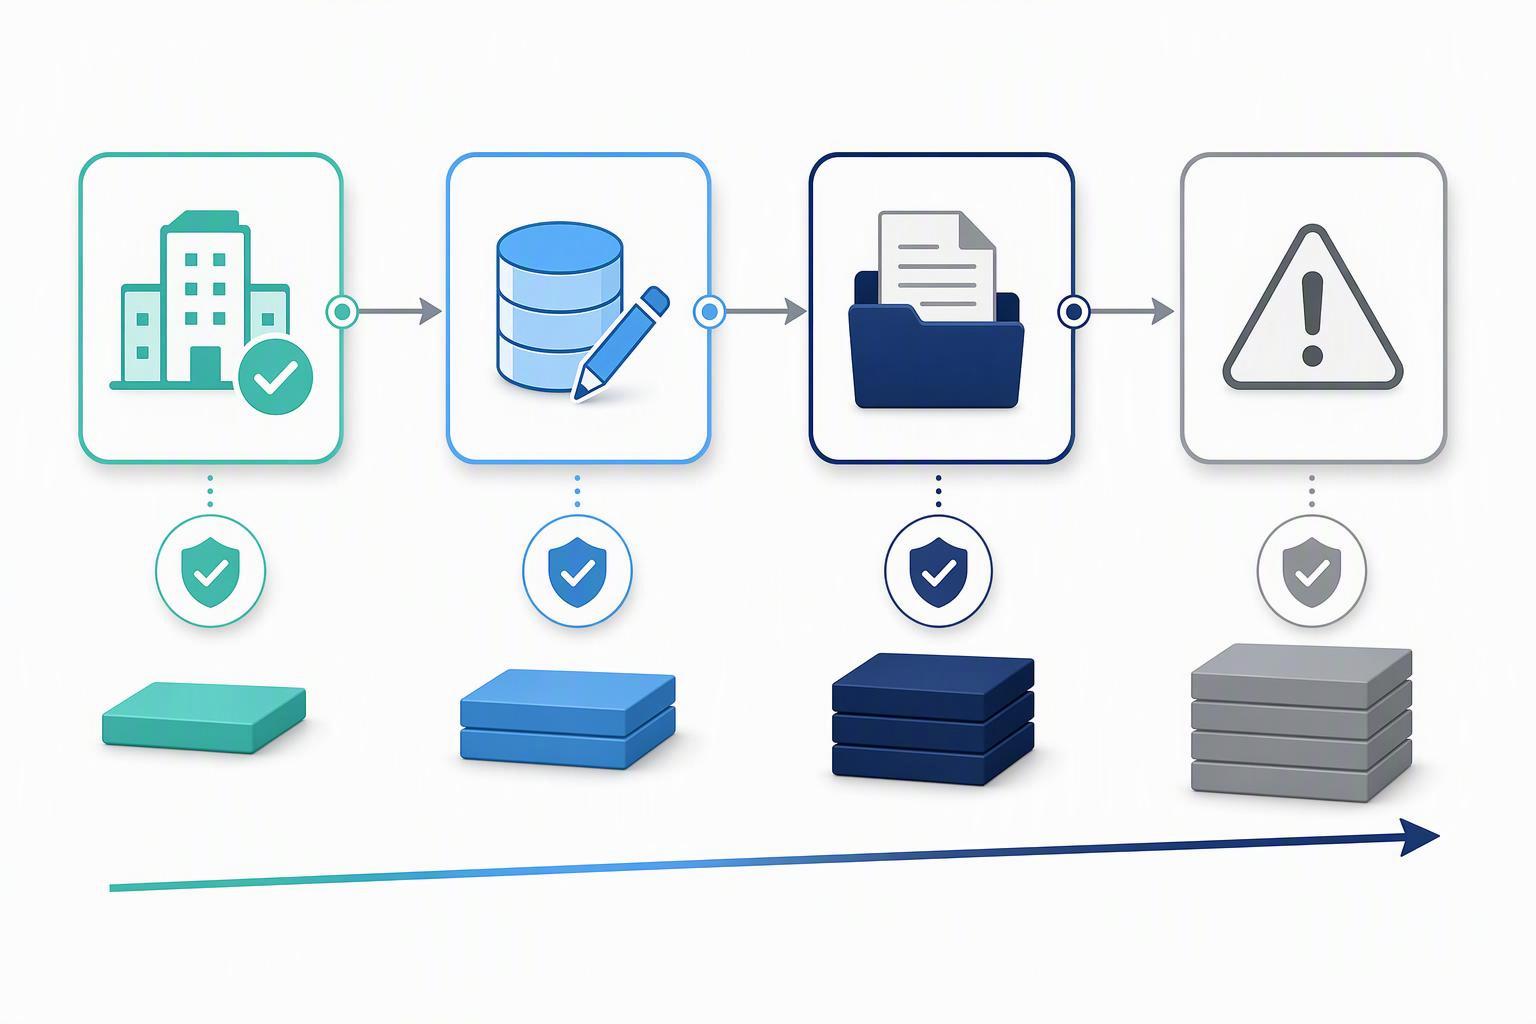
User fees fund the review system; they do not buy approval. That distinction is the whole reason peptide supplier qualification now has to be built as an evidence file rather than a vendor scorecard.
PDUFA’s reauthorization through September 2027 keeps the prescription drug user-fee program running on a five-year cycle, FY 2023 through FY 2027, under the FDA User Fee Reauthorization Act of 2022 signed on 30 September 2022 (FDA, Prescription Drug User Fee Amendments, retrieved 28 August 2026). What that money does is narrower than it sounds. In the FDA’s own explanation of its user-fee programs, the fees “supplement, not replace” congressional appropriations and are “not a ‘fee-for-service’ payment,” and review outcomes do not depend on whether a fee was collected (FDA, FDA: User Fees Explained, retrieved 22 Mei 2026). The same page is explicit that performance goals are not guarantees: FDA may keep working past a review goal date, and the goals “do not count on meeting the timeline 100% of the time” (FDA, FDA: User Fees Explained, retrieved 22 Mei 2026).
Enforcement runs on the same machinery, and it does not wait for a fee cycle. In the May 2026 warning letter that put a peptide API supplier on Import Alert 66-40, FDA found that the quality unit failed to ensure CGMP compliance and failed to maintain complete traceability of APIs in commercial distribution; the cited product was semaglutide and tirzepatide API, not peptides generically, but the finding is about the record, not the molecule (FDA Warning Letter 723330, 1 Mei 2026).
The failure pattern repeats in composite: a shipment sits at the border because the file cannot show who actually made the material. Registration and listing, entry data, entry documents on request, detention risk. The burden accumulates at each stage, and no stage accepts a supplier’s assurance in place of a record.
Step 1: Define What You Must Be Able to Prove Before You Vet Anyone
Your qualification file exists to answer four questions: who made the material, under what quality system, how it is identified, and who is accountable for it at the border. Everything you collect later is evidence for one of those four answers.
The regulatory baseline explains why the bar sits where it does. FDA states that active pharmaceutical ingredients are inherently misbranded under FD&C Act 502(f)(1) because their labeling lacks adequate directions for consumer use, and that they must comply with 21 CFR 201.122 to be eligible for a labeling exemption (FDA, Importing Active Pharmaceutical Ingredients, retrieved 2026-06-15). The same FDA guidance notes that bulk drug substances used in compounding must comply with either section 503A or 503B of the FD&C Act, and that foreign establishments which manufacture, repack, relabel or salvage APIs imported into the United States must register with FDA under FD&C Act 510(i) and 21 CFR 207.17, listing all known importers in that registration.
Sintesis Peptida Those three requirements map directly onto the four questions. Accountability at the border is not a character judgment about your vendor; it is a registration and listing status you can check.
Note: This framework is an evaluation method, not a legal opinion. Obligations vary by jurisdiction and by the material’s intended use, so confirm requirements with your own regulatory and legal counsel.
Step 2: Qualify the Vendor Against Licensure, cGMP and Inspection History
A supplier’s registration status and inspection history are checkable facts, not assurances. The buyer who skips the check inherits the finding.
Start with licensure and listing. Commercially distributed APIs must be listed with FDA under section 510(j) and 21 CFR 207.41, and at the time of importation that listing must be the drug-listing requirement that must be the manufacturer’s own, not a distributor’s. Then verify cGMP status and inspection history directly, and confirm the vendor sits on your approved supplier list with a documented requalification trigger.
Skipping this step has a documented cost. FDA’s the supplier-vetting failures FDA cited in December 2025 letter to Darmerica LLC found the quality unit failed to ensure API CGMP compliance, requalified a supplier with inconsistent inspectional history and no stability data, and failed to list distributed drugs. That case involves API supply generally rather than peptides specifically, but the qualification logic transfers directly.
|
Claim type |
Document that Peptida Sintetik satisfies it |
What its absence signals |
|---|---|---|
|
Licensure and registration |
Foreign establishment registration and importer listing |
Unverified legal basis to import |
|
Drug listing |
Manufacturer’s own API listing under 510(j) |
Distributor listing masking the true manufacturer |
|
cGMP status |
Current compliance status and applicable guidance |
Reliance on self-declaration |
|
Inspection history |
Inspection records and any warning letter history |
Unassessed enforcement risk |
|
Approved supplier list |
Controlled list with requalification criteria |
No change control over the supplier base |
The failure mode is substitution. FDA’s the batch-number and date changes FDA documented in the 2026 warning letter to Harbin Jixianglong Biotech describes semaglutide API repackaged and relabeled from suppliers not on the approved supplier list, issued new batch numbers such as CP-030-20250711, with the manufacturer misidentified and manufacturing and retest dates altered without supporting data. A CGMP finding at that point converts into Import Alert 66-40, the detention-without-physical-examination mechanism, which holds shipments automatically.
Step 3: Handle Sequence Information as a Controlled Input
Sequence information is a controlled input with its own screening and retention obligations, and a supplier that cannot describe its escalation rule in writing has not implemented one.
The federal screening framework’s screening thresholds start at DNA or RNA 200 nucleotides or longer, with the length recommended to drop to 50 nt within three years, and the best-match approach defined over a 66 amino acid / 200 nt window (HHS Screening Framework, 88 FR 2023-22540, 2023-10-13). The amino-acid best-match extension in the screening framework applies that same window across all six reading frames, so a peptide order is not outside the screening conversation just because it arrives as an amino acid sequence.
The guidance on orders below the screening threshold recommends screening short orders where components could be assembled into larger sequences of concern, and encourages benchtop synthesizer manufacturers to build screening into equipment (HHS ASPR Screening Framework FAQ, updated 2025-11-19). Ask what happens when a hit occurs: who reviews it, against what rule, and what the customer is asked to supply. The list of documents that can establish a customer’s legitimate use includes proposed end use, institutional affiliation, a named biosafety officer, internal review records, FSAP registration or a completed BIS-711, publication history, ORCID, business licenses, grant numbers, or a research plan. The federal screening framework’s record-retention expectations set a floor of at least three years for sequences-of-concern orders, rising to eight where not unduly burdensome.
Step 4: Build Material Traceability From Order Intake to Third-Party Transfer
A Certificate of Analysis is one node in the trail, not the endpoint. The trail has to hold when the material changes hands, and that is where most files break.
Under ALCOA+, analytical records must be attributable, legible, contemporaneous, original, accurate, complete, consistent, enduring and available. Applied to peptides, that test reaches past the CoA into the raw data behind it: HPLC chromatograms, MS spectra, integration parameters, deviations and failed injections. ICH Q2(R2)’s validation expectations for the methods behind a CoA set the method standard; ALCOA+ is the data-integrity framing regulators now apply to the records those methods generate.
Perkhidmatan The documented failure pattern is quieter than a missing document. It is a new batch number created over existing material, a manufacturer misidentified on a transfer record, dates changed without supporting data, the same traceability failure FDA cited in the 2026 warning letter.
Key Takeaway: A lot number identifies material. A batch record explains it. Buyers auditing peptide supply chain traceability need both, linked.
Step 5: Prepare Export and Shipping Documentation That Clears the First Review
The document set is requested, not volunteered, so assemble it before the shipment moves. Any FDA-regulated product offered for import must meet the same standards as domestic products, and incomplete or invalid electronic data routes a shipment to human review under FDA’s ImportShield Program (FDA, Entry Review, retrieved 2026-01-21).
That review can ask for copies of product labels, certificates, ingredient lists, processing records, and analytical information, submitted through ITACS, FDA’s preferred channel for entry-review documents. The source lists document categories rather than a mandated universal set, so treat the list as the ceiling of what may be requested, not a fixed checklist.
In practice, your peptide export and shipping documentation file should hold a commercial invoice whose line descriptions match the entry data exactly, a packing list, and an SDS. For lyophilized peptides, add the temperature-control record covering the transit window.
Step 6: Assemble the Vendor Qualification Documentation Package
The vendor qualification documentation package is not a folder of PDFs. It is a set of documents with a named owner and a review date for Shop each item, so that regulatory, QA, legal and IP reviewers can each find what they need without asking you to reconstruct the file.
|
Package item |
Primary reviewer |
Typical retention |
|---|---|---|
|
Licensure and registration evidence |
Regulatory |
Life of the supplier relationship |
|
cGMP and inspection history |
QA |
Life of the supplier relationship |
|
Sequence-handling and access records |
IP / legal |
Per your records policy |
|
Lot-level Certificate of Analysis and traceability records |
QA |
Per your records policy |
|
Export and shipping documentation set |
Regulatory / legal |
Per customs and tax rules in each jurisdiction |
Assign the owner before you circulate the package. A document nobody owns is a document nobody updates.
A representative example of how the pieces map to a supplier’s own published materials: MOL Changes issues a Certificate of Analysis per batch, with lot numbers searchable on its CoA page, alongside per-production-run identity, purity and quantitation testing by HPLC coupled with mass spectrometry. Whether that structure fits your file is a judgement for your reviewers, not a claim about outcomes.
Confirm retention periods against your own regulatory and legal obligations; they vary by jurisdiction and by material.
Common Mistakes That Sink a Qualification File
The most common failure is not a missing document. It is a document that cannot be traced to the material in the box. Five patterns account for most of the files that collapse under review.
Treating the CoA as the whole trail. A certificate proves what a batch tested at, not where it went. The traceability failures documented in the 2026 warning letter turned on exactly this gap, plus date changes that no receiving record could corroborate. Fix: pair every CoA with intake records, transfer records and the dates each was signed.
Accepting a registration claim without checking the listing. Buyers take a vendor’s word that its facility is registered. The requirement that the listing be the manufacturer’s own is what reviewers check first. Fix: pull the listing yourself and confirm the entity name matches the invoice.
Having no written sequence-screening escalation rule. Screening gets done informally, so nobody can show what happened when a hit appeared. The screening framework’s escalation expectations are procedural, not discretionary. Fix: write the threshold, the reviewer and the timeline before the first order.
Shipping on a generic commercial invoice. Descriptions like “research compound” invite the questions that what FDA entry review can ask you to produce is designed to answer. Fix: describe the material as the CoA does. Pengeluaran Peptida
Requalifying a supplier with inconsistent inspectional history. This is the requalification failure FDA cited in December 2025: no stability data, no resolution of prior findings. Fix: make requalification conditional on closed findings, not elapsed time.
Frequently Asked Questions
Can I rely on a distributor’s documentation instead of qualifying the manufacturer?
Tidak. Registration and listing obligations attach to the manufacturer, and the listing presented at the time of importation must be the manufacturer’s own API listing. A distributor’s paperwork describes a commercial relationship, not the facility that made the material, so it does not substitute for qualifying the manufacturer directly. Ask the distributor to identify the manufacturing site, then verify that site’s registration, listing and inspection record yourself.
What happens if my shipment is routed to human review?
FDA’s ImportShield screening flags entries for review, and a flagged entry can be referred to the district for a documentation request. Expect to supply the entry documents, the manufacturer’s listing evidence, and product-specific records on request. ITACS is the preferred channel for submitting that response, and the clock runs from the referral, so keep the document set assembled before the shipment moves rather than after it is held. Tentang
How long do I need to keep sequence-screening and order records?
The Federal Register recommendation is at least 3 years for sequence-of-concern orders, and 8 years where retention is not unduly burdensome. The IGSC protocol sets an 8-year member standard, which is the stricter of the two. If your own counsel has not set a period, 8 years keeps you aligned with both.
Conclusion
The five pillars form one framework: define what you must prove, qualify the vendor against licensure and inspection history, treat sequence information as a controlled input, trace material from intake to third-party transfer, and ship documentation that clears first review. The deliverable is a defensible internal case, not a vendor recommendation. A peptide supplier qualification file earns its keep when your regulatory, legal, and IP reviewers can follow every claim back to a primary document.
If you are weighing a specific sequence, the practical next step is a technical feasibility assessment: send the sequence, target scale, purity requirement, and destination market, and ask for the documentation package you would need to qualify the supplier. MOL Changes is a specialized R&D organization integrating organic chemistry and biology, offering custom peptide synthesis from sequence design to large-scale production with HPLC, MS, and sterility quality control.
Disclosure: MOL Changes is a custom peptide synthesis provider, so we have a commercial interest in this topic. Regulatory obligations vary by jurisdiction; confirm your specific requirements with your own regulatory and legal counsel before relying on any framework described here.