सबूत का बोझ पेप्टाइड आपूर्तिकर्ता योग्यता पर क्यों चला गया
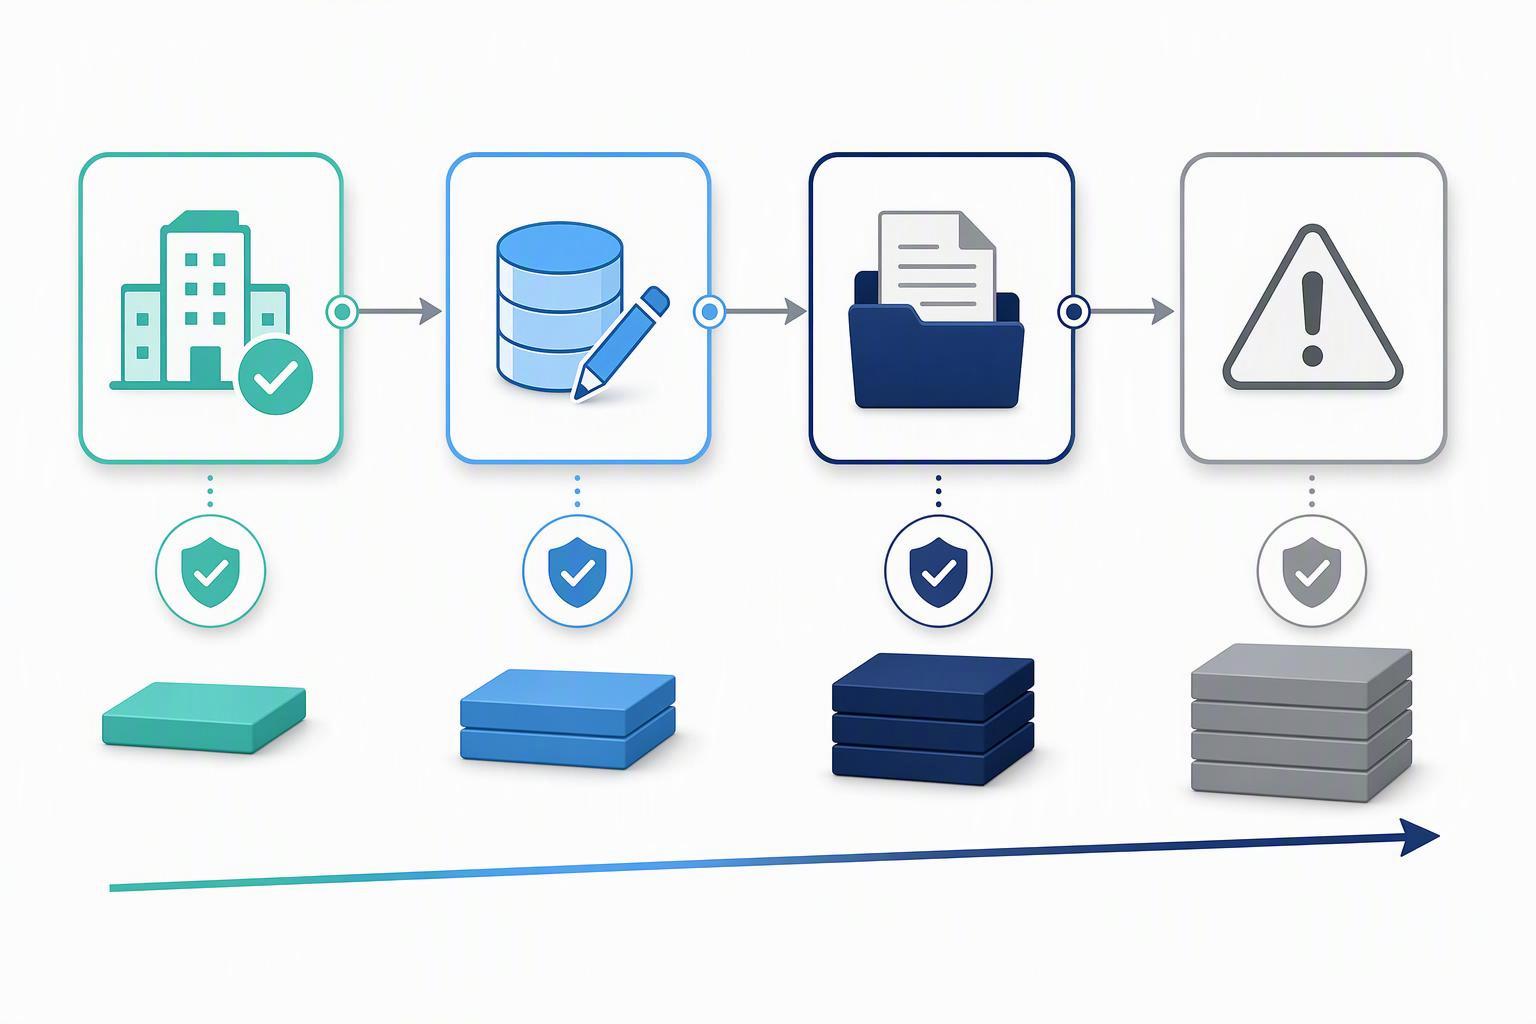
उपयोगकर्ता शुल्क समीक्षा प्रणाली को निधि देता है; वे अनुमोदन नहीं खरीदते हैं. यही अंतर है जिसके कारण पेप्टाइड आपूर्तिकर्ता योग्यता को अब विक्रेता स्कोरकार्ड के बजाय एक साक्ष्य फ़ाइल के रूप में बनाया जाना है.
सितंबर तक पीडीयूएफए का पुन:प्राधिकरण 2027 प्रिस्क्रिप्शन ड्रग उपयोगकर्ता-शुल्क कार्यक्रम को पांच साल के चक्र पर चालू रखता है, वित्तीय वर्ष 2023 वित्तीय वर्ष के माध्यम से 2027, एफडीए उपयोगकर्ता शुल्क पुनर्प्राधिकरण अधिनियम के तहत 2022 पर हस्ताक्षर किये गये 30 सितम्बर 2022 (एफडीए, प्रिस्क्रिप्शन ड्रग उपयोगकर्ता शुल्क संशोधन, पुनर्प्राप्त 28 अगस्त 2026). वह पैसा जो करता है वह जितना लगता है उससे कहीं अधिक संकीर्ण है. अपने उपयोगकर्ता-शुल्क कार्यक्रमों के बारे में FDA की अपनी व्याख्या में, फीस “पूरक।”, कांग्रेस के विनियोगों को प्रतिस्थापित नहीं करते हैं और ये 'सेवा के लिए शुल्क' भुगतान नहीं हैं,” और समीक्षा के नतीजे इस बात पर निर्भर नहीं करते कि शुल्क एकत्र किया गया था या नहीं (एफडीए, एफडीए: उपयोगकर्ता शुल्क समझाया गया, पुनर्प्राप्त 22 मई 2026). वही पृष्ठ स्पष्ट है कि प्रदर्शन लक्ष्य गारंटी नहीं हैं: एफडीए समीक्षा लक्ष्य तिथि के बाद भी काम करना जारी रख सकता है, और लक्ष्य “समयसीमा को पूरा करने पर निर्भर नहीं हैं।” 100% समय का" (एफडीए, एफडीए: उपयोगकर्ता शुल्क समझाया गया, पुनर्प्राप्त 22 मई 2026).
प्रवर्तन उसी मशीनरी पर चलता है, और यह शुल्क चक्र की प्रतीक्षा नहीं करता है. मई में 2026 चेतावनी पत्र जिसने पेप्टाइड एपीआई आपूर्तिकर्ता को आयात चेतावनी पर डाल दिया 66-40, एफडीए ने पाया कि गुणवत्ता इकाई सीजीएमपी अनुपालन सुनिश्चित करने में विफल रही और वाणिज्यिक वितरण में एपीआई की पूर्ण पता लगाने की क्षमता बनाए रखने में विफल रही; उद्धृत उत्पाद सेमाग्लूटाइड और टिरजेपेटाइड एपीआई था, सामान्यतः पेप्टाइड्स नहीं, लेकिन खोज रिकॉर्ड के बारे में है, अणु नहीं (एफडीए चेतावनी पत्र 723330, 1 मई 2026).
विफलता पैटर्न समग्र रूप से दोहराता है: एक शिपमेंट सीमा पर खड़ा है क्योंकि फ़ाइल यह नहीं दिखा सकती कि वास्तव में सामग्री किसने बनाई है. पंजीकरण एवं सूचीकरण, प्रवेश डेटा, अनुरोध पर प्रवेश दस्तावेज़, हिरासत का जोखिम. प्रत्येक चरण पर बोझ बढ़ता जाता है, और कोई भी स्तर रिकॉर्ड के स्थान पर आपूर्तिकर्ता के आश्वासन को स्वीकार नहीं करता है.
कदम 1: परिभाषित करें कि किसी की जांच करने से पहले आपको क्या साबित करने में सक्षम होना चाहिए
आपकी योग्यता फ़ाइल चार प्रश्नों के उत्तर देने के लिए मौजूद है: सामग्री किसने बनाई, किस गुणवत्ता प्रणाली के तहत, इसकी पहचान कैसे की जाती है, और सीमा पर इसके लिए कौन जिम्मेदार है. बाद में आप जो कुछ भी एकत्र करेंगे वह उन चार उत्तरों में से एक का प्रमाण होगा.
विनियामक आधार रेखा बताती है कि बार वहीं क्यों बैठता है जहां वह बैठता है. एफडीए का कहना है कि एफडी के तहत सक्रिय फार्मास्युटिकल सामग्री स्वाभाविक रूप से गलत ब्रांडेड हैं&सी एक्ट 502(एफ)(1) क्योंकि उनके लेबलिंग में उपभोक्ता उपयोग के लिए पर्याप्त दिशा-निर्देशों का अभाव है, और इसका उन्हें अनुपालन करना होगा 21 सीएफआर 201.122 लेबलिंग छूट के लिए पात्र होना (एफडीए, सक्रिय फार्मास्युटिकल सामग्री आयात करना, पुनर्प्राप्त 2026-06-15). वही एफडीए मार्गदर्शन नोट करता है कि कंपाउंडिंग में उपयोग किए जाने वाले थोक दवा पदार्थों को एफडी की धारा 503ए या 503बी का अनुपालन करना चाहिए।&सी एक्ट, और वह विदेशी प्रतिष्ठान जो निर्माण करते हैं, हिन्दी repack, संयुक्त राज्य अमेरिका में आयातित रीलेबल या सेल्वेज एपीआई को एफडी के तहत एफडीए के साथ पंजीकृत होना चाहिए&सी एक्ट 510(मैं) और 21 सीएफआर 207.17, उस पंजीकरण में सभी ज्ञात आयातकों को सूचीबद्ध करना.
पेप्टाइड संश्लेषण वे तीन आवश्यकताएँ सीधे चार प्रश्नों पर आधारित होती हैं. सीमा पर जवाबदेही आपके विक्रेता के चरित्र का निर्णय नहीं है; यह एक पंजीकरण और लिस्टिंग स्थिति है जिसे आप जांच सकते हैं.
टिप्पणी: यह रूपरेखा एक मूल्यांकन पद्धति है, कानूनी राय नहीं. दायित्व क्षेत्राधिकार और सामग्री के इच्छित उपयोग के अनुसार भिन्न-भिन्न होते हैं, इसलिए अपने स्वयं के नियामक और कानूनी परामर्शदाता से आवश्यकताओं की पुष्टि करें.
कदम 2: लाइसेंस के विरुद्ध विक्रेता को अर्हता प्राप्त करें, सीजीएमपी और निरीक्षण इतिहास
आपूर्तिकर्ता की पंजीकरण स्थिति और निरीक्षण इतिहास जांच योग्य तथ्य हैं, आश्वासन नहीं. जो खरीदार चेक छोड़ देता है उसे परिणाम विरासत में मिलता है.
लाइसेंस और लिस्टिंग से शुरुआत करें. व्यावसायिक रूप से वितरित एपीआई को अनुभाग के तहत एफडीए के साथ सूचीबद्ध किया जाना चाहिए 510(जे) और 21 सीएफआर 207.41, और आयात के समय वह सूचीकरण दवा-सूचीकरण आवश्यकता होनी चाहिए जो निर्माता की अपनी होनी चाहिए, वितरक का नहीं. फिर सीधे सीजीएमपी स्थिति और निरीक्षण इतिहास सत्यापित करें, और दस्तावेजी पुनर्योग्यता ट्रिगर के साथ पुष्टि करें कि विक्रेता आपकी स्वीकृत आपूर्तिकर्ता सूची में है.
इस चरण को छोड़ने पर एक दस्तावेजी लागत आती है. एफ.डी.ए एफडीए ने दिसंबर में आपूर्तिकर्ता-परीक्षण विफलताओं का हवाला दिया 2025 डार्मरिका एलएलसी को लिखे पत्र में पाया गया कि गुणवत्ता इकाई एपीआई सीजीएमपी अनुपालन सुनिश्चित करने में विफल रही, असंगत निरीक्षण इतिहास और बिना स्थिरता डेटा वाले आपूर्तिकर्ता को पुन: योग्य घोषित कर दिया गया, और वितरित दवाओं को सूचीबद्ध करने में विफल रहा. उस मामले में विशेष रूप से पेप्टाइड्स के बजाय आम तौर पर एपीआई आपूर्ति शामिल है, लेकिन योग्यता तर्क सीधे स्थानांतरित हो जाता है.
|
दावा प्रकार |
उसका दस्तावेजीकरण करें सिंथेटिक पेप्टाइड्स इसे संतुष्ट करता है |
इसकी अनुपस्थिति क्या संकेत देती है |
|---|---|---|
|
लाइसेंस और पंजीकरण |
विदेशी प्रतिष्ठान पंजीकरण और आयातक सूचीकरण |
आयात करने का असत्यापित कानूनी आधार |
|
औषधि सूची |
नीचे निर्माता की अपनी एपीआई सूची है 510(जे) |
वितरक सूची असली निर्माता को छुपा रही है |
|
सीजीएमपी स्थिति |
वर्तमान अनुपालन स्थिति और लागू मार्गदर्शन |
स्व-घोषणा पर निर्भरता |
|
निरीक्षण इतिहास |
निरीक्षण रिकॉर्ड और किसी चेतावनी पत्र का इतिहास |
मूल्यांकन न किया गया प्रवर्तन जोखिम |
|
अनुमोदित आपूर्तिकर्ता सूची |
पुनः योग्यता मानदंड के साथ नियंत्रित सूची |
आपूर्तिकर्ता आधार पर कोई परिवर्तन नियंत्रण नहीं |
विफलता मोड प्रतिस्थापन है. एफडीए के बैच-नंबर और तारीख में बदलाव को एफडीए ने दस्तावेज में दर्ज किया है 2026 हार्बिन जिक्सियांग्लोंग बायोटेक को लिखे चेतावनी पत्र में सेमाग्लूटाइड एपीआई को उन आपूर्तिकर्ताओं से पुनः पैक और पुनः लेबल किए जाने का वर्णन किया गया है जो अनुमोदित आपूर्तिकर्ता सूची में नहीं हैं।, सीपी जैसे नए बैच नंबर जारी किए-030-20250711, निर्माता की गलत पहचान की गई और सहायक डेटा के बिना निर्माण और पुन: परीक्षण की तारीखें बदल दी गईं. उस बिंदु पर एक सीजीएमपी खोज आयात चेतावनी में परिवर्तित हो जाती है 66-40, हिरासत-बिना-शारीरिक-परीक्षा तंत्र, जो शिपमेंट को स्वचालित रूप से रखता है.
कदम 3: अनुक्रम जानकारी को नियंत्रित इनपुट के रूप में संभालें
अनुक्रम जानकारी अपने स्वयं के स्क्रीनिंग और प्रतिधारण दायित्वों के साथ एक नियंत्रित इनपुट है, और एक आपूर्तिकर्ता जो लिखित रूप में अपने वृद्धि नियम का वर्णन नहीं कर सकता है, उसने इसे लागू नहीं किया है.
संघीय स्क्रीनिंग ढांचे की स्क्रीनिंग सीमाएँ डीएनए या आरएनए से शुरू होती हैं 200 न्यूक्लियोटाइड या उससे अधिक, जिस लंबाई तक छोड़ने की अनुशंसा की गई है 50 तीन साल के भीतर नहीं, और सर्वोत्तम-मिलान दृष्टिकोण को परिभाषित किया गया है 66 एमिनो एसिड / 200 एनटी विंडो (एचएचएस स्क्रीनिंग फ्रेमवर्क, 88 फादर 2023-22540, 2023-10-13). स्क्रीनिंग फ्रेमवर्क में अमीनो-एसिड बेस्ट-मैच एक्सटेंशन सभी छह रीडिंग फ़्रेमों पर एक ही विंडो लागू करता है, इसलिए पेप्टाइड ऑर्डर केवल इसलिए स्क्रीनिंग वार्तालाप से बाहर नहीं है क्योंकि यह अमीनो एसिड अनुक्रम के रूप में आता है.
स्क्रीनिंग सीमा से नीचे के आदेशों पर मार्गदर्शन छोटे आदेशों की स्क्रीनिंग की सिफारिश करता है जहां घटकों को चिंता के बड़े अनुक्रमों में इकट्ठा किया जा सकता है, और बेंचटॉप सिंथेसाइज़र निर्माताओं को उपकरण में स्क्रीनिंग बनाने के लिए प्रोत्साहित करता है (एचएचएस एएसपीआर स्क्रीनिंग फ्रेमवर्क अक्सर पूछे जाने वाले प्रश्न, अद्यतन 2025-11-19). पूछें कि जब कोई हिट होती है तो क्या होता है: जो इसकी समीक्षा करता है, किस नियम के विरुद्ध, और ग्राहक को क्या आपूर्ति करने के लिए कहा गया है. दस्तावेज़ों की सूची जो ग्राहक के वैध उपयोग को स्थापित कर सकती है, उसमें प्रस्तावित अंतिम उपयोग भी शामिल है, संस्थागत संबद्धता, एक नामित जैव सुरक्षा अधिकारी, आंतरिक समीक्षा रिकॉर्ड, एफएसएपी पंजीकरण या पूर्ण बीआईएस-711, प्रकाशन इतिहास, ORCID, व्यापार लाइसेंस, अनुदान संख्या, या एक शोध योजना. संघीय स्क्रीनिंग ढाँचे की रिकॉर्ड-प्रतिधारण अपेक्षाएँ चिंता के क्रम के आदेशों के लिए कम से कम तीन साल की सीमा निर्धारित करती हैं, जहां यह अनावश्यक रूप से बोझिल नहीं है, आठ तक बढ़ रहा है.
कदम 4: ऑर्डर लेने से लेकर तीसरे पक्ष के स्थानांतरण तक सामग्री का पता लगाने की क्षमता बनाएं
विश्लेषण का प्रमाणपत्र पथ में एक नोड है, समापन बिंदु नहीं. जब सामग्री हाथ बदलती है तो निशान को पकड़ना पड़ता है, और यहीं पर अधिकांश फ़ाइलें टूट जाती हैं.
ALCOA+ के तहत, विश्लेषणात्मक रिकॉर्ड जिम्मेदार होने चाहिए, पढ़ने योग्य, समसामयिक, मूल, शुद्ध, पूरा, सुसंगत, स्थायी और उपलब्ध. पेप्टाइड्स पर लागू, वह परीक्षण सीओए से आगे निकलकर उसके पीछे के कच्चे डेटा तक पहुंचता है: एचपीएलसी क्रोमैटोग्राम, एमएस स्पेक्ट्रा, integration parameters, deviations and failed injections. मैं Q2(आर2)’s validation expectations for the methods behind a CoA set the method standard; ALCOA+ is the data-integrity framing regulators now apply to the records those methods generate.
सेवाएं The documented failure pattern is quieter than a missing document. It is a new batch number created over existing material, a manufacturer misidentified on a transfer record, dates changed without supporting data, the same traceability failure FDA cited in the 2026 warning letter.
कुंजी ले जाएं: A lot number identifies material. A batch record explains it. Buyers auditing peptide supply chain traceability need both, linked.
कदम 5: निर्यात और शिपिंग दस्तावेज़ तैयार करें जो पहली समीक्षा को स्पष्ट करता हो
The document set is requested, not volunteered, so assemble it before the shipment moves. Any FDA-regulated product offered for import must meet the same standards as domestic products, and incomplete or invalid electronic data routes a shipment to human review under FDA’s ImportShield Program (एफडीए, Entry Review, पुनर्प्राप्त 2026-01-21).
That review can ask for copies of product labels, प्रमाण पत्र, ingredient lists, processing records, and analytical information, submitted through ITACS, FDA’s preferred channel for entry-review documents. The source lists document categories rather than a mandated universal set, so treat the list as the ceiling of what may be requested, not a fixed checklist.
व्यवहार में, your peptide export and shipping documentation file should hold a commercial invoice whose line descriptions match the entry data exactly, a packing list, and an SDS. For lyophilized peptides, add the temperature-control record covering the transit window.
कदम 6: विक्रेता योग्यता दस्तावेज़ीकरण पैकेज को इकट्ठा करें
The vendor qualification documentation package is not a folder of PDFs. It is a set of documents with a named owner and a review date for दुकान each item, so that regulatory, क्यूए, legal and IP reviewers can each find what they need without asking you to reconstruct the file.
|
Package item |
Primary reviewer |
Typical retention |
|---|---|---|
|
Licensure and registration evidence |
नियामक |
Life of the supplier relationship |
|
cGMP and inspection history |
क्यूए |
Life of the supplier relationship |
|
Sequence-handling and access records |
आई पी / कानूनी |
Per your records policy |
|
Lot-level Certificate of Analysis and traceability records |
क्यूए |
Per your records policy |
|
Export and shipping documentation set |
नियामक / कानूनी |
Per customs and tax rules in each jurisdiction |
Assign the owner before you circulate the package. A document nobody owns is a document nobody updates.
A representative example of how the pieces map to a supplier’s own published materials: MOL Changes issues a Certificate of Analysis per batch, with lot numbers searchable on its CoA page, alongside per-production-run identity, purity and quantitation testing by HPLC coupled with mass spectrometry. Whether that structure fits your file is a judgement for your reviewers, not a claim about outcomes.
Confirm retention periods against your own regulatory and legal obligations; they vary by jurisdiction and by material.
सामान्य गलतियाँ जो योग्यता फ़ाइल को डुबा देती हैं
The most common failure is not a missing document. It is a document that cannot be traced to the material in the box. Five patterns account for most of the files that collapse under review.
Treating the CoA as the whole trail. A certificate proves what a batch tested at, not where it went. The traceability failures documented in the 2026 warning letter turned on exactly this gap, plus date changes that no receiving record could corroborate. Fix: pair every CoA with intake records, transfer records and the dates each was signed.
Accepting a registration claim without checking the listing. Buyers take a vendor’s word that its facility is registered. The requirement that the listing be the manufacturer’s own is what reviewers check first. Fix: pull the listing yourself and confirm the entity name matches the invoice.
Having no written sequence-screening escalation rule. Screening gets done informally, so nobody can show what happened when a hit appeared. The screening framework’s escalation expectations are procedural, not discretionary. Fix: write the threshold, the reviewer and the timeline before the first order.
Shipping on a generic commercial invoice. Descriptions like “research compound” invite the questions that what FDA entry review can ask you to produce is designed to answer. Fix: describe the material as the CoA does. पेप्टाइड उत्पादन
Requalifying a supplier with inconsistent inspectional history. This is the requalification failure FDA cited in December 2025: no stability data, no resolution of prior findings. Fix: make requalification conditional on closed findings, not elapsed time.
अक्सर पूछे जाने वाले प्रश्नों
Can I rely on a distributor’s documentation instead of qualifying the manufacturer?
नहीं. Registration and listing obligations attach to the manufacturer, and the listing presented at the time of importation must be the manufacturer’s own API listing. A distributor’s paperwork describes a commercial relationship, not the facility that made the material, so it does not substitute for qualifying the manufacturer directly. Ask the distributor to identify the manufacturing site, then verify that site’s registration, listing and inspection record yourself.
यदि मेरा शिपमेंट मानव समीक्षा के लिए भेजा जाता है तो क्या होगा??
FDA’s ImportShield screening flags entries for review, and a flagged entry can be referred to the district for a documentation request. Expect to supply the entry documents, the manufacturer’s listing evidence, and product-specific records on request. ITACS is the preferred channel for submitting that response, and the clock runs from the referral, so keep the document set assembled before the shipment moves rather than after it is held. के बारे में
मुझे अनुक्रम-स्क्रीनिंग और ऑर्डर रिकॉर्ड कितने समय तक रखने की आवश्यकता है??
The Federal Register recommendation is at least 3 years for sequence-of-concern orders, और 8 years where retention is not unduly burdensome. The IGSC protocol sets an 8-year member standard, which is the stricter of the two. If your own counsel has not set a period, 8 years keeps you aligned with both.
निष्कर्ष
The five pillars form one framework: define what you must prove, qualify the vendor against licensure and inspection history, treat sequence information as a controlled input, trace material from intake to third-party transfer, and ship documentation that clears first review. The deliverable is a defensible internal case, not a vendor recommendation. A peptide supplier qualification file earns its keep when your regulatory, कानूनी, and IP reviewers can follow every claim back to a primary document.
If you are weighing a specific sequence, the practical next step is a technical feasibility assessment: send the sequence, target scale, purity requirement, and destination market, and ask for the documentation package you would need to qualify the supplier. एमओएल चेंजेस एक विशेष आर है&डी संगठन कार्बनिक रसायन विज्ञान और जीव विज्ञान को एकीकृत करता है, offering custom peptide synthesis from sequence design to large-scale production with HPLC, एमएस, and sterility quality control.
खुलासा: MOL Changes is a custom peptide synthesis provider, so we have a commercial interest in this topic. Regulatory obligations vary by jurisdiction; confirm your specific requirements with your own regulatory and legal counsel before relying on any framework described here.