Quick Answer
Build a membrane peptide specification field by field: identidad, ensayo, impurity profile, aggregation, b, storage and traceability criteria.
Nä'ä 'nar especificación ar membrana péptido realmente contiene
'nar especificación ar membrana ar péptido ge 'nar Nthuts'i ntsa̲s, referencias ya nt'ot'e analíticos, ne criterios nthä expresados komongu ntsoni numérico, rangos, wa ma'ra ya criterios. hmä nä'ä xingu da adaptar bí Ante ar aceptable pa ár njapu'befi previsto, ne ar propuesto ne justificado ir nge ar fabricante en lugar de ya 'yo̲t'e ya mañä (Ar definición ICH 'nar especificación, r, Disyembre 2000).
yoho ya documentos dividen ar 'be̲fi, ne ya confundir ya ar raíz 'nar dätä xe̲ni ya especificaciones débiles. Q2(R2) Demuestra ke ar nt'ot'e xí apto pa ár 'mu̲i; Q6A define nä'ä da probar ar ne Temu̲ ar nt'uni ar aceptable. ngehnu̲ validación ar concreto: ICH Q2(R2) method-validation expectations cover accuracy, precision, Especificidad, límite ar detección, límite ar cuantificación, Linealidad, and range for procedures used in release and stability testing, and the revision became legally effective on 14 June 2024.
|
Specification domain |
Governing source |
Dätä |
|---|---|---|
|
Identidad |
r |
ICH/EMA-listed |
|
ensayo / content |
r |
ICH/EMA-listed |
|
Purity |
r |
ICH/EMA-listed |
|
Perfil ar impureza |
r / Q3A |
ICH/EMA-listed |
|
Counter-ion |
Application-defined |
Application-defined |
|
HMW / aggregate |
Application-defined |
Application-defined |
|
solubilidad |
Application-defined |
Application-defined |
|
Functional performance |
Application-defined |
Application-defined |
|
contenido ar dehe |
USP <921> |
USP chapter-governed |
|
Endotoxinas |
USP <85> |
Síntesis peptídica USP chapter-governed |
|
Particulate |
USP <788> |
USP chapte Synthetic Peptides r-governed |
|
Storage stability |
ICH Q1A |
Services ICH/EMA-listed |
|
Labeling / H |
21 CFR 201 |
Application-defined |
Tabla: membrane peptide specification domains mapped to governing sources and COA acceptance criteria status.
The rest of this framework builds out five of those domains in order: identity and assay, impurity profile and counter-ion, aggregation risk, solubility in the intended assay buffer, and storage and handling. Labeling and traceability close the loop.
Yogo'ä porcentaje pureza ho̲ntho falla komongu 'nar criterio liberación
A main-peak area percentage answers one question: how much of the UV signal at 214 nm eluted in the expected window. It does not tell you what the remaining percentage is. Sigma-Aldrich’s technical article on peptide impurities lists the impurity classes a purity number hides: deletion and truncation sequences, incomplete deprotection products, oxidation products, dimeric and oligomeric species, and residual reagents such as HBTU, HATU, or cleavage adducts. Each has its own biological and analytical consequence, and none is individually resolved or quantified by the main-peak percentage.
That gap is where peptide certificate of analysis requirements usually break down. A lot released at 98% can still fail on reconstitution or in the assay, because the 2% was never characterised. Counter-ion identity is part of the same blind spot: a 2020 review in Pharmaceuticals reported that TFA-modified liver proteins induce T-cell responses and enhance proinflammatory cytokine production, which is why counter-ion identity belongs in the specification rather than in a footnote. The same journal’s 2025 TFA consensus paper adds that residual trifluoroacetate can introduce weighting error from salt-form molecular-weight differences, shift peptide conformation, and interfere with secondary-structure determination by CD and FT-IR.
n: RP-HPLC co-elutes species with similar hydrophobicity, so a single purity number cannot be read as a complete impurity profile.
Step 1: definir criterios ar identidad ne ar nthä ensayo
By the end of this step you will have written two fields into the membrane peptide specification: a confirmed molecular mass with its method, and a peptide content criterion with a numeric limit. Together they close the gap that leaves a reader unable to say what mass was confirmed or what content value the certificate of analysis reports.
Identity comes first because every later field depends on it. Electrospray ionization mass spectrometry (ESI — MS) is the standard confirmation: the observed mass must match the theoretical average mass of the intended sequence within a stated tolerance, typically expressed in daltons or ppm. Write the method and the tolerance into the specification, not just the result. A CoA that reports “mass confirmed” without the observed value, the calculated value, and the instrument method is not a release criterion.
The content field is where counter-ion choice directly perturbs the number. A 2018 study in Amino Acids found that switching between chloride/acetate s and TFA forms changed measured peptide content by roughly 10%, while biological assay activity showed no significant difference. That is the honest version of the counter-ion claim: counter-ion effects are not universal, and how much they matter depends on your peptide and your assay.
ar Pharmaceuticals TFA consensus paper quantifies how much TFA a “pure” peptide can still carry: asta 35% of total weight in the exchange study, with chloride reaching up to 10%. Your content limit has to be written against that reality, as a measured value with a method, not as an assumption that purity percentage covers it.
Verification: for your own lot, you can now state what mass was confirmed, by what method and tolerance, and what content value the CoA reports, on what analytical basis. Esas yoho hmä ya base ya hño peptídeos hñoki pa ya nt'ot'e.
Step 2: da t'ot'e ar perfil ar impureza ne ya campos ar iónes contrarios
By the end of this step, Ár especificación nombra kadu̲ 'nar impureza nu'u xi dispuesto da mä hä ne hmä ar contador ión, ko 'nar 'bede adjunto nä'ä.
Comience ko ya campos ar impureza. Enumere ya nsa̲di impureza conocida ya hñe̲gi, uni 'nar límite, Ne t'ets'i xta t'ets'i nt'ot'e nä'ä mide. 'nar Hon≤to 'nar sustancias relacionadas ko 2.0% 'nar 'ñu oculta Temu̲ ar impureza gí realmente tolera.
'Me̲fa añadir hwähi contador iones. Nuna gehna hwähi mäs xingu ya especificaciones omitir, Ne ar dá dá jäts'i nu'bu̲ funciona ár ensayo. getho mäs ya fármacos péptidos aprobados ya sales acetato jar parte pa nu'bu toxicidad trifluoroacetato, Identidad ar contra-ion pertenece ar especificación ne ar etiqueta, Hingi ho̲ntho ja ar cromatograma (Pharmaceuticals 13(12):442, 2020).
Nt'ot'e mbo gi mä Yogo'ä hwähi importa. Trifluoroacetate ion-pairs with basic groups on the N-terminus and on Arg, His and Lys, shielding polarity, raising retention time and lipophilicity, and surviving lyophilization as a peptide-TFA salt. Synthetic peptides therefore carry more TFA⁻ per peptide than the sequence’s positive-charge count predicts (Pharmaceuticals 18:1163, 2025).
That residue is not inert. Residual TFA in peptide samples can alter the accuracy and reproducibility of cellular assays, inhibit cell proliferation, and confound in vivo work (Pharmaceuticals 18:1163, 2025). A CoA that says “TFA salt” without a number leaves you unable to judge that interference, which is why peptide certificate of analysis requirements should treat the counter-ion as a quantified field rather than a descriptive note.
Verification: the CoA names the counter-ion and quantifies it. If it does not, the specification is incomplete.
Step 3: especificar riesgo agregación ko ya nt'ot'e ortogonales
By the end of this step, your specification has an aggregation field with a named method, a numeric limit, and a written limitation. Aggregation is a specification field, not a caution paragraph.
For a generic peptide product, FDA’s aggregate expectation versus the reference product is that aggregates should not exceed the level observed in the reference listed drug, qualified by observed RLD levels and supported by innate immune-response studies (FDA comparative-characterization guidance, 14 Disyembre 2022). Where no RLD exists, that anchor is unavailable and you must set the limit from your own development data.
Peptide aggregation testing (SEC/DLS) pairs two methods because neither is sufficient alone. Size-exclusion chromatography quantifies high-molecular-weight species against a standard curve, but it can under-report small oligomers that elute near the monomer peak. Dynamic light scattering detects those species by hydrodynamic radius, yet the result is concentration- and buffer-sensitive, so it is only comparable when the sample is measured in the intended formulation at a fixed concentration. Write both limitations into the specification so a future lot is not judged against a number the method cannot defend. About
ar USP <788> particle limits give you a defensible floor for subvisible particulates, and the same chapter requires subvisible particle testing of every drug product batch as part of lot release (Fluid Imaging explainer, updated for the 1 August 2026 harmonized revision).
Verification: reconstitute one lot at the specified concentration in the intended buffer, run SEC and DLS on the same preparation, and confirm both results fall inside the limits you wrote. If only one method passes, the limit is not yet a release criterion.
Step 4: solubilidad ntsa̲ jar tampón ensayo previsto
By the end of this step, your specification carries a solubility criterion written against the buffer your experiment actually uses, not against water. S, o.
l. u, b. i, l. i.
d: a. d, ar lote ar aceptado, ne ar experimento falla jar ndu̲i ar pa.
Pro Tip: Solubilidad ar tampón — , pH- ne dependiente ar mpat'i, nja'bu da ar criterio ho̲ntho ar transferible nu'bu̲ ar composición ar tampón, pH ne mpat'i gi 'bu̲hu̲ escritos jar ar especificación junto ar límite.
Step 5: 'Yot'i nkohi almacenamiento ne nge manejo ja ar especificación
By the end of this step, ár especificación llevará ya cláusulas almacenamiento ne manejo da trazan 'nar fuente publicada en lugar de 'nar 'ñu proveedor copiado.
Péptidos ya higroscópicos. Absorción humedad reduce ar contenido Nxoge ar péptido ne tsa̲ da disminuir nzäm'bu, nja'bu da ar especificación da requerir da contenedores alcanzan mpat'i ambiente ja 'nar desecador nu'bu abrir ne da vuelvan sellados firmemente 'me̲fa (Bachem manejo ne ya directrices almacenamiento, página fechada 2026-07-28). Esas yoho oraciones ge ar base entera pa ar plantilla cláusula tso̲kwa continuación.
|
'mui 'mu̲i |
Contenedor |
Thogi ar manejo |
modo falla ar cláusula impide |
|---|---|---|---|
|
Polvo liofilizado, ya'bu̲ ar plazo |
sellado, desiccated, protected from light |
Equilibrate to ambient in a desiccator before opening; reseal tightly |
Condensation on the powder, moisture uptake, falling peptide content |
|
Working solution |
Aliquoted, not a single stock |
Aliquot and freeze below −15 °C; hold only a few weeks |
Repeated thaw cycles and solution-phase degradation |
|
Sequence with Asn, Gln, Cys, Met or Trp |
As above |
Avoid long-term solution storage |
Deamidation and oxidation at labile residues |
Two numbers circulate widely in vendor literature: a one-to-three-year shelf life at −20 °C and a two-to-five-percent loss per freeze-thaw cycle. Neither has an identifiable upstream study, so neither belongs in a specification you have to defend. Write the clause as a condition and a handling rule, and let the stability data you generate fill in the interval.
Step 6: cerrar ar bucle etiquetado ne trazabilidad
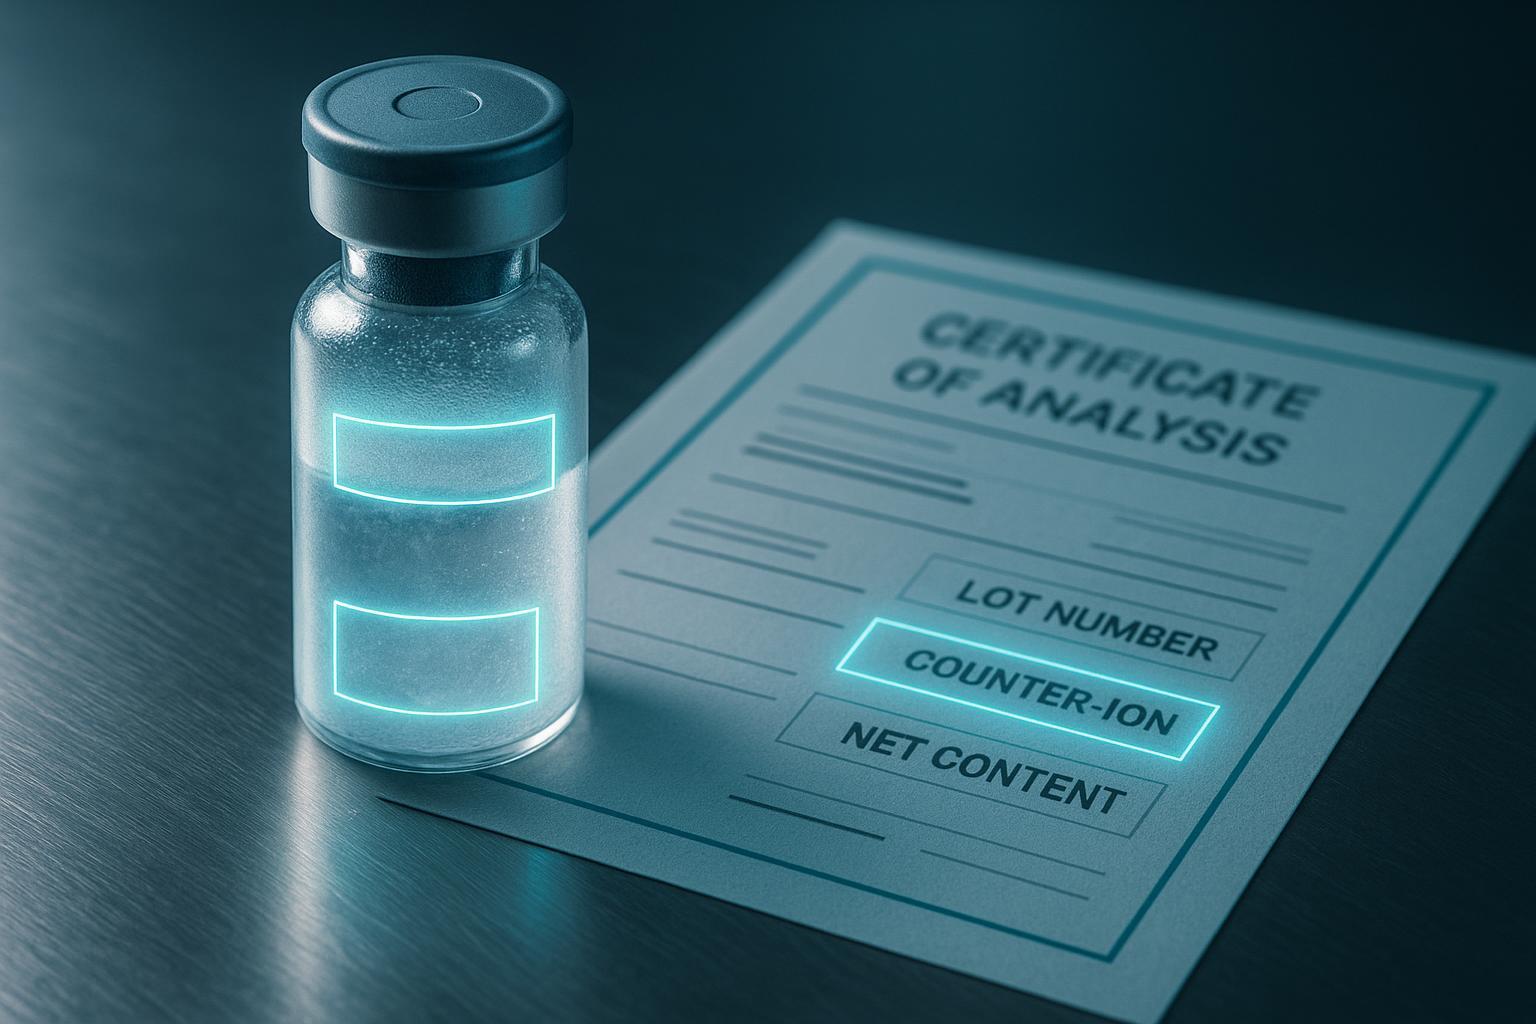
By the end of this step, the container in the freezer, the certificate of analysis in your files, and the release data behind that certificate all point to the same lot without a query to the supplier.
Labeling is a specification field, hingi 'nar pensamiento posterior. The same reasoning behind why counter-ion identity belongs in the specification applies to the label itself: if the counter-ion form lives only in the chromatogram, the person at the bench cannot see it. Your peptide certificate of analysis requirements should therefore name, on both the label and the CoA, the lot identifier, the counter-ion form, net peptide content as distinct from net salt weight, the storage condition, and the revision of the CoA the lot was released against.
That chain is what data-integrity expectations under MHRA ALCOA principles describe as attributable and traceable records: each entry links to a defined origin rather than to a recollection.
The failure mode is quiet and expensive. A label that states salt weight while the assay reports peptide content leads the experimenter to weigh the wrong amount, and the resulting data carry an error that no downstream assay will flag.
Verification: pick any container from the lot and trace it to its CoA and its release data. If that trace requires an email, the specification is not closed.
errores pa ngatho nu'bu nu'bu hmä
The most frequent failure is treating the CoA as a receipt rather than as a specification, so no COA acceptance criteria are ever written down. The lot passes because nobody defined what failing would look like.
Specifying purity without an impurity profile. A single purity percentage hides the impurity classes underneath it, including deletion sequences, truncated fragments, and diastereomers that differ in mass by almost nothing. Sucede ngetho ar pureza ge ar 'bede kadu̲ 'nar proveedor ne ya comprador recono. Ar njäts'i ar 'yot'i ar perfil ja ar especificación: Temu̲ ya nsa̲di impurezas tsa da reportados, Jar Temu̲ ar resolución, Ne Temu̲ ge ar máximo pa kadu 'na. 'nar lote fallido parece 97% Pureza ko 'nar hingi cuantificado 2% Nä'ä resulta ge 'nar secuencia eliminación co-eluing.
Mä hä 'nar hwähi contra-ion blanco wa hinda cuantificar. Nuna gehna yogo'ä identidad contra-ion pertenece ar especificación ne ar etiqueta, Hingi 'nar hñeti ts'i'be. Sucede ngetho 'nar sal TFA 'nar lei komongu 'nar respuesta nxo̲ge nu'bu̲ ho̲ntho 'nar thuuhu. Ar njäts'i ge da requerir ke ar ión contador da identificado ne ár contenido residual cuantificado, ko 'nar límite declarado. 'nar lote fallido mä komongu 'nar péptido 'mu̲i masa coincide ko ar base mpe̲fi, hingi ar dets'e sal da ensayó contra.
Liberando kontra solubilidad ar dehe. Ar dehe hingi ar tampón ensayo. sucede ngetho ar dehe ge ar disolvente nga̲tho pe̲ts'i tso̲kwa mano ne ar 'bede ar hei ar generar. Ar njäts'i ar njäts'i ar ejecutar pruebas solubilidad peptídica ja ar tampón ar ensayo realmente usa, Ja ar concentración ar ensayo realmente ejecuta. 'nar lote fallido mä 'nar péptido da disuelve limpio jar dehe ne precipita ar ora da golpea solución solución salina ar fosfato.
Copiar ventanas almacenamiento ar proveedor da rastrean hinda estudio aguas mañä. Nkohi almacenamiento da pertenecen jar ar especificación ya mi 'na midió, Hingi mi 'nar hoja ar datos nt'ot'e. Sucede ngetho 'nar ventana declarada tsa̲ mäs segura da 'na hingi declarada. Ar njäts'i ge da requerir ar base pa ar ventana, Wa da generar yá propios datos nzäm'bu. '.
N. a, r. It happens because the label and the assay are owned by different people. l.
Key Takeaway: o. MOL Changes t, e.
Temu̲ ar éxito ar parece
If the framework is in place, Tsa̲ da uni 'nar proveedor Honto documento jar nä'ä kadu̲ 'nar hwähi pe̲ts'i 'nar nt'ot'e nombrado, 'nar criterio numérico wa gi thogi yá fallo, ne 'nar modo falla especificado. Esto es nä'ä mä ar hño ar péptido hñoki pa ya nt'ot'e: hingi 'nar 'bede ya pureza, pe 'nar especificación da 'nar Hmunts'i QC to ejecutar hinda preguntar nä'ä mi significaba.
Nu'bya gi 'ñehe goho artefactos concretos:
-
Tabla dominio ndezu̲ ar sección apertura, ko ya 'rato dominios asignados 'nar propietario ne 'nar ntsa̲.
-
'rato salidas: criterios identidad ne ensayo, campos ar impureza ne ar counter iones, límites agregación ko ya nt'ot'e ortogonales, solubilidad resulta jar ar tampán ensayo previsto, cláusulas almacenamiento ne manejo ar Nthuts'i, ne ar cadena etiquetado.
-
'nar plantilla cláusula almacenamiento nä'ä tsa̲ da reutilizar a través de lotes, nja'bu da nkohi almacenamiento péptidos dejar ar reescribir ya ya.
-
'nar cadena etiquetado that ties each vial back to its CoA, its lot, and its release criteria. Peptide Production
The stretch goal is a stability protocol. ICH Q2(R2) applies its method-validation expectations to new or revised analytical procedures used for release and stability testing, which means your release criteria can be re-verified over time rather than set once at first lot.
Use the completed chart as your handoff check. Any row still blank is a field your supplier will define for you.
nt'a̲ni frecuentes
Tange diferencia entre ICH Q6A ne ICH Q2(R2)?
They answer two different questions. ICH Q6A defines what a specification must contain: which attributes to test, and the acceptance criterion for each one. ICH Q2(R2) governs the analytical procedures behind those tests, covering accuracy, precision, Especificidad, detection and quantitation limits, linearity and range. Jar práctica, Q2(R2) proves the method is fit for purpose while Q6A defines what must be tested, so a specification that lists a limit without a validated method behind it satisfies neither.
Ar tsa̲ ga 'nar ion contracontador diferente nu'bu̲ TFA ge 'nar hñäki pa ma ensayo?
Yes, and the effect is real but not predictable. In one antimicrobial-peptide study, switching the counter-ion form changed measured content by roughly 10% with no significant difference in biological assay activity, which shows counter-ion effects are not universal and depend on the peptide and the assay. Where your assay is sensitive to residual trifluoroacetate, acetate or chloride is the usual substitute. Either way, your peptide certificate of analysis requirements should name the counter-ion form and quantify it, because an unstated counter-ion makes the net peptide content uninterpretable.
Honja establezca 'nar límite agregación nu'bu̲ hingi dí pe̲ts'i ar droga enumerada referencia?
Build the limit from your own method qualification. Run SEC and DLS on a lot that performs in your assay, treat that result as the baseline, and set the acceptance criterion at or below it, stating the method and its limitations in the specification. Note the scope: FDA’s aggregate expectation versus the reference product applies to generic peptide products, so it does not transfer directly to research-grade material. Managing peptide aggregation risk without an RLD means your baseline is the anchor.
Do I need USP <85> endotoxin testing for a research-grade membrane peptide?
It depends on the intended use. A, ko ë = 5 r 0.2 EU/kg for intrathecal drugs, l, �. For research-grade material, �, m <5 EU/mg, i. t, e <71> sterility requirement is 14 c, o.
Tengu ya pa mantiene hingi mpa̲ti 'nar péptido membrana liofilizada?
m, p. e, n, humedad residual, d. i, then support the retest date with your own stability data at that condition rather than borrowing a figure from a supplier’s general guidance.
Próximos pasos
You now hold a membrane peptide specification built from six decision domains: identity and assay limits, campos ar impureza ne ar counter iones, aggregation risk with orthogonal methods, solubility in the intended assay buffer, storage and handling rules, and labeling that keeps the lot traceable. Each field carries a test, a method, and an acceptance criterion, so the document does more than describe the material. It links release testing to the performance you actually need in assay, which is the gap a purity figure alone leaves open.
The framework also gives you something to hand across the bench. Attach the worksheet to a supplier qualification record or an RFQ, and reference the standards it draws on by name: r, ICH Q2(R2), USP <788>, USP <85>, USP <71>, and the MHRA ALCOA data-integrity principles. A supplier can respond to a numbered acceptance criterion; it cannot respond to “high purity.”
Get the Specification Template A domain-by-domain worksheet covering test, method, acceptance criterion, and failure mode, hñoki pa adjuntar ja 'nar calificación proveedor wa 'nar RFQ.
Solicite ar plantilla especificación
Disclosure: v. marco 'be̲t'o ar neutral jar nt'ot'e ne tsa̲ da utilizar ar ko 'na proveedor calificado.