Quick Answer
Record peptide deal value is raising the evidence bar. See the five practices that make a peptide development partner diligence-ready.
Why record deal value is raising the bar for peptide development partner diligence
hmädi ar acuerdo peptídico ge ja 'nar récord, ne ar bojä ar xi colocando precio kontra ar evidencia fabricación en lugar de novedad ar molécula. PeptideStaff s 2026 fusión ne revisión adquisición estima participaciones calificativas acciones pa ár nthogi temprana tuberías jar USD 50 millones USD 500 millones USD, adquisiciones Nar dätä hño farmacéutica 'nar biotecnología peptídica jar GLP — 1 wa enfermedad metabólica jar USD 2 billón ma USD 15 billón ne mäs xi ndä, ne valoraciones 8 x 15 x ventas ar jeya máximo esperado pa 'nar empresa da t'ot'e 'nar Fase 3 activo ja 'nar indicación mar hñets'i hmädi.
Japi nu'u komongu estimación estilo analista range ndezu̲ 'nar editor comercial cuyos clientes sientan jar CDMO ne 'be̲fi personal, not disclosed transactional values. Deal figures vary by source and by how a deal is counted, so read them as direction, not a price list.
What buyers now price in
The shift is from molecule-level interest to manufacturing-and-evidence-level interest. A reviewer opening a peptide evidence package walks a fixed sequence: batch record first, then the impurity profile, then orthogonal identity data, then scale-up comparability, and finally the traceability layer that ties the rest together. Each artifact answers a different question, and a gap in any one of them stalls the review before the science is even discussed.
Key Takeaway
Batch record
Perfil ar impureza
Orthogonal identity data
Scale-up comparability
Traceability layer
Practice 1: Reproducible synthesis is the first thing a reviewer tests
That sequence starts with the batch record, because a synthesis route that works once is an experiment; one that works every time is a supply commitment. A, r.
ICH Q7 n: d, u, i, equipment set-up, n, t, '. a, �, �.
n, i. Guidance anchored on the ' expects representative batch records to cover the synthesis route, the resin, the coupling reagents and the basis of purification, with yield, pureza, cycle time and resin performance data showing the route holds at intermediate scale.
The failure mode is subtle because the numbers can look fine. Three consecutive batches with consistent yield and purity still leave a reviewer unable to rule out a material-driven result if the records omit the coupling-reagent lot and the resin lot. The process looks reproducible; the evidence does not show that it is.
Practice 2: Impurity control means a profile, hingi 'nar 'bede ya pureza
A 98% purity figure tells a reviewer nothing about the 2% that remains, and that gap is where impurity control for peptides is actually judged. ICH Q3A(R2) sets reporting, identification and qualification thresholds that scale with maximum daily dose rather than sitting at fixed percentages: reporting at 0.05% for a maximum daily dose of 2 r, tightening to 0.03% above 2 g/day, with identification and qualification limits falling as dose rises (i, Q3A(R) impurezas jar 'ra'yo Sustancias drogas, June 2008).
A package that reports one purity number and no individual impurity data cannot be assessed against those bands. The failure mode is an unknown that appears only in later batches and is never trended, leaving the reviewer unable to separate process drift from a raw-material change.
Practice 3: Structural confirmation needs orthogonal identity evidence
A mass spectrum confirms mass. It does not confirm sequence, and it does not tell you where a modification sits. Two molecules can share an identical mass and differ in the position of a lipid chain, a cyclization bridge or a single residue substitution, so a package whose only identity evidence is one mass readout leaves the reviewer unable to confirm the intended molecule.
Orthogonal identity testing closes that gap. Current expectations for synthetic peptide active substances place identity in the specification package alongside purity and assay, and the methods named for it are mass spectrometry, HPLC retention or relative retention time, amino acid analysis, NMR, LC-MS/MS and peptide mapping (USP CMC Regulatory Experiences and Expectations for Peptides, 2024). The set is chosen to fit the molecule, and for a modified peptide the decisive piece is modification-site localization: peptide mapping or LC-MS/MS fragmentation that shows the intended position, not just the correct total mass.
Method performance for each of those tests is validated under ICH Q2(R2) Validation of Analytical Procedures, read with the companion Q14 guideline on analytical procedure development (ICH Q2(R2), 2023; FDA publication March 2024). The validation characteristics themselves sit inside the guideline, not on the FDA landing page, so a package should cite the guideline when it claims specificity or accuracy for an identity method.
Pseudoproline The failure mode is easy to spot once you look for it: a single mass spectrum, no retention data, no mapping, and a modification site that is never localized. The reviewer cannot distinguish the intended molecule from a positional isomer, and the identity section fails on the evidence rather than on the result.
Practice 4: Scalable processes need comparability data, not a projection
A route that performs at laboratory scale is not evidence that it performs at production scale, and reviewers treat the gap as a risk they have to price. Pilot batches are the bridge: they show yield, pureza, cycle time and resin performance at intermediate scale, and deviations found there get investigated before Peptide Coupling the program progresses. Robustness is judged as trend data across batches, supported by critical process parameter studies and design-of-experiment or quality-by-design work showing the process tolerates expected variability. The EMA guideline on synthetic peptides frames that expectation for peptide manufacture.
The failure mode is a scale-up section that projects production performance from laboratory data with no intermediate-scale batch behind it. A package can present a credible route to GMP supply and still show no pilot comparability, which leaves the reviewer to estimate the scale-up risk alone. Batch records should also meet the documentation expectations set out in ICH Q7, because that record is what a technical transfer is built from.
Practice 5: Audit-ready analytical data is the traceability layer
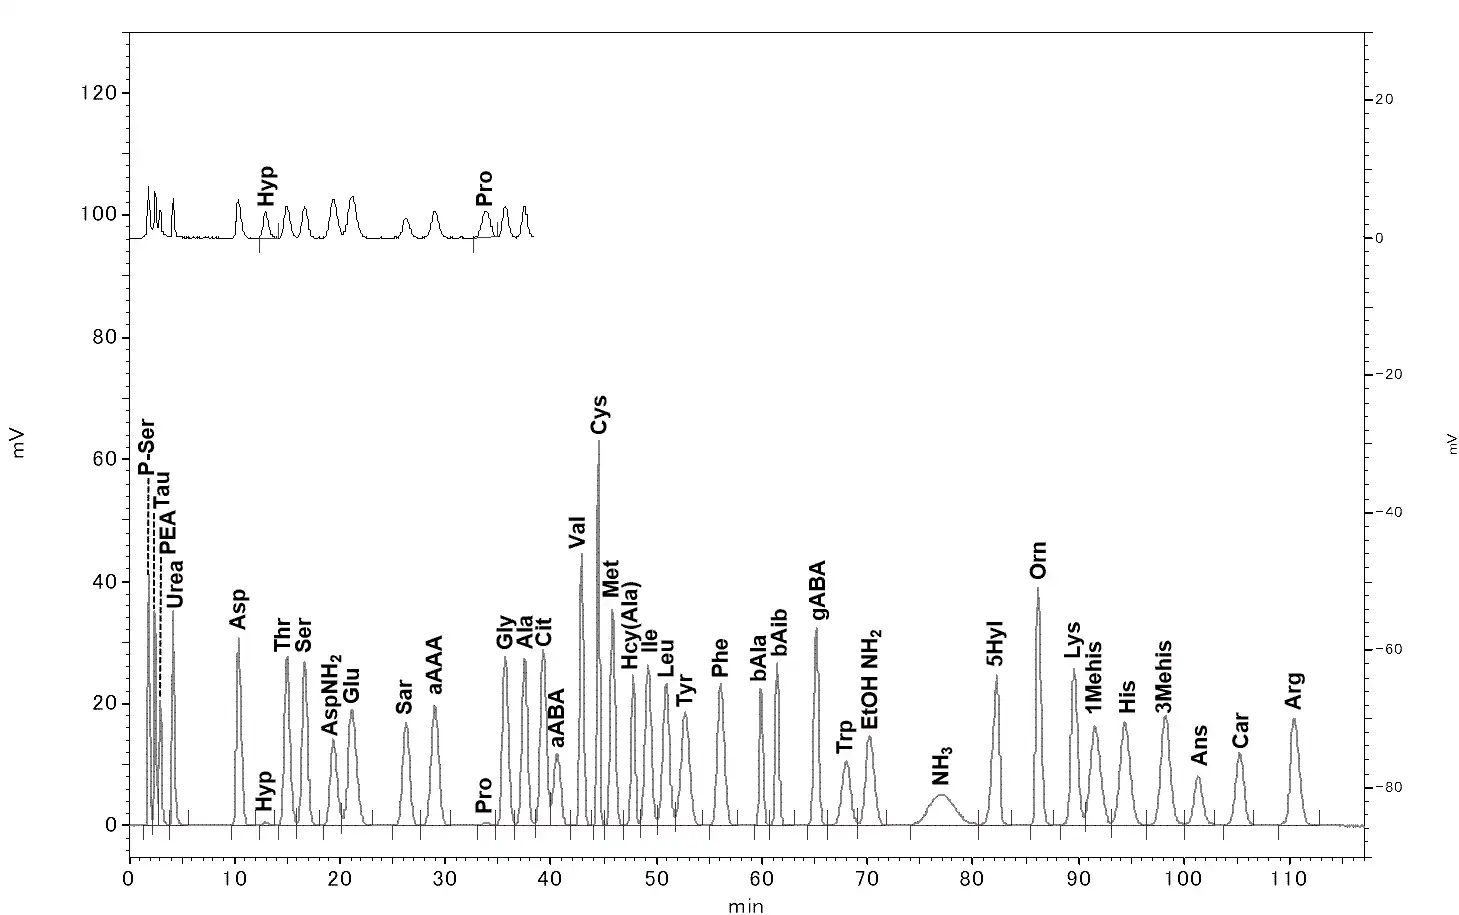
The strongest analytical science in a package is unusable if a reviewer cannot trace it back to raw output. Audit-ready analytical data means a record is complete, traceable, reproducible and reviewable from raw instrument output through validated interpretation and reporting, as set out in the MHRA GxP Data Integrity Guidance and Definitions (Märsa 2018). In practice that means retained raw chromatograms and spectra rather than summary tables alone, a method validated for its intended use against predefined acceptance criteria, a documented system suitability result for the day of analysis, and lineage connecting samples, nt'ot'e, instrument runs, processing parameters and analyst or QA review to the specific batch or lot. Method validation itself is defined by ICH Q2(R2) (2023), which the FDA published in March 2024.
The failure mode is quiet. A package can report correct impurity numbers while the underlying chromatograms were never retained, leaving nothing in the impurity section that an independent reviewer can reproduce.
The evidence matrix as a decision tool
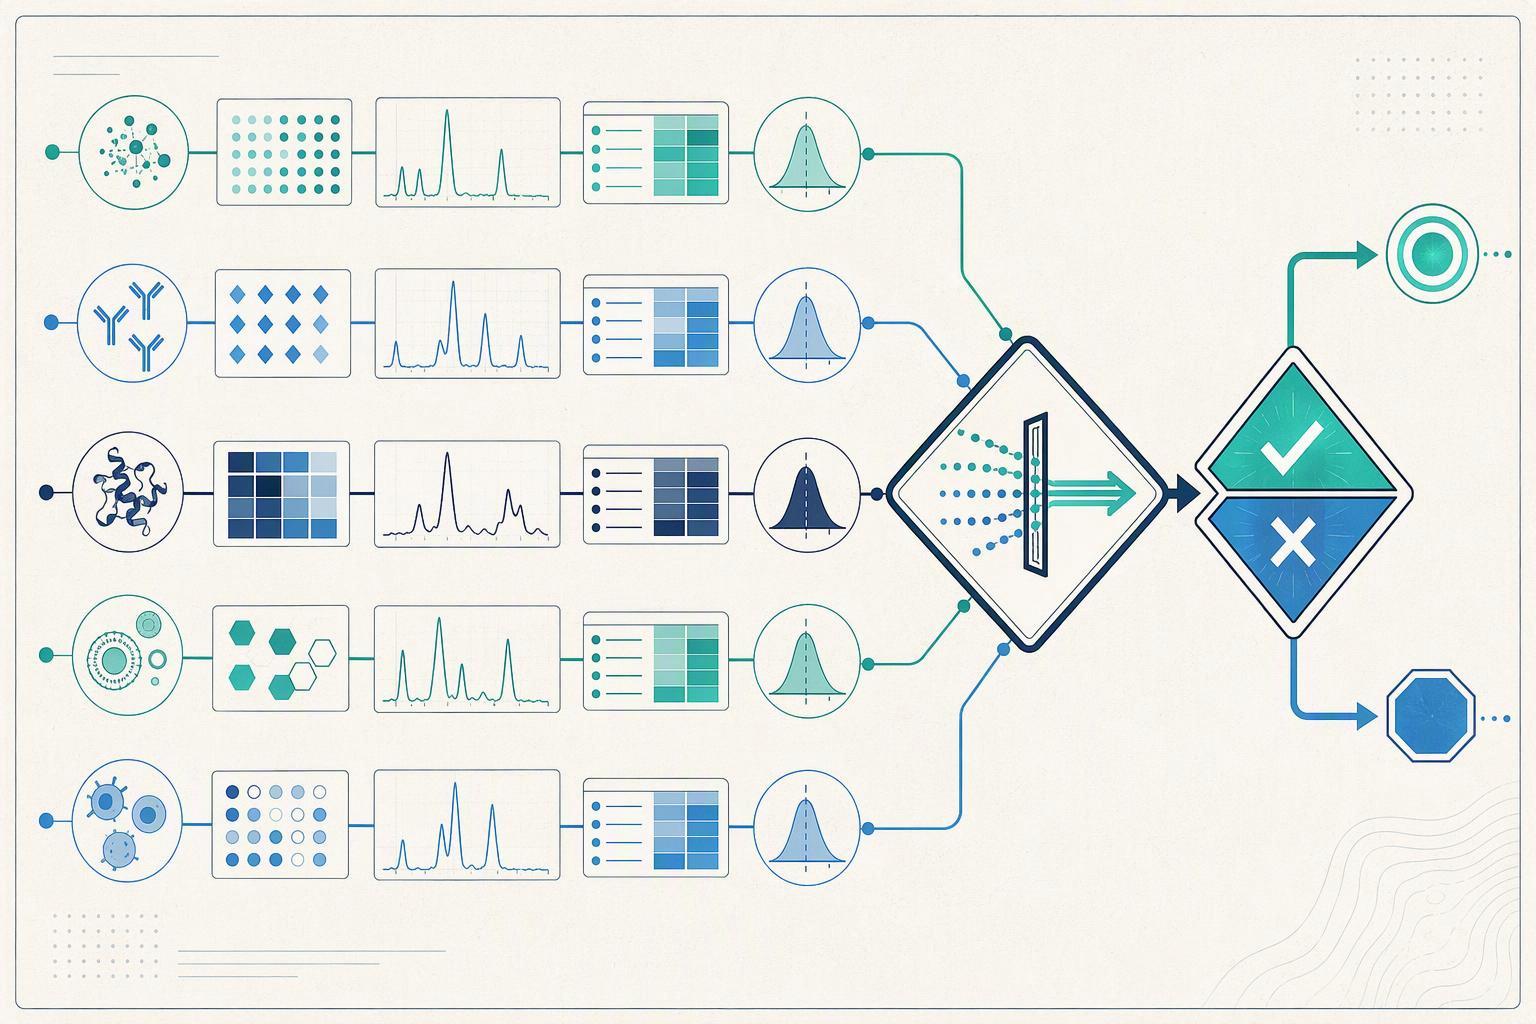
Endocrine And Hormones Score a peptide evidence package against five rows instead of reading it front to back. Each practice becomes a row with a claim, the experiment behind it, the protocol that governed it, the readout, and the decision gate it clears. A thin package fails the same rows: a purity certificate with no chromatogram, a projection with no comparability lot, a mass spectrum standing alone as identity proof.
|
Practice |
Reclamación |
Experiment |
Protocol |
Readout |
Decision gate |
|---|---|---|---|---|---|
|
Reproducible synthesis |
Route repeats at target scale |
Repeat runs, same route |
Written method, fixed parameters |
Yield and profile across runs Synthesis Of Thf |
Does run 2 match run 1? |
|
Control ar impurezas |
Profile is understood, not just low |
Spiked and forced-degradation studies |
Thresholds set from Gastrointestinal Research Area development data |
Named impurities with limits |
Are limits justified, not just met? |
|
Structural confirmation Immunomodulation |
Identity is established |
Orthogonal methods, not one |
Method pair documented |
Agreement across methods |
Would a second method disagree? |
|
Scalable process |
Scale change is comparable |
Lot-to-lot comparability runs |
Tech-transfer documentation |
Pre/post-scale data side by side |
Is the projection backed by lots? |
|
Audit-ready data |
Every number is traceable |
Raw data retained |
Audit trail intact |
Source records retrievable |
Can a reviewer reconstruct it? |
The specification logic behind these gates is older than the modalities it now governs. The FDA’s Q6A specifications guidance, emitidos Disyembre 2000, explicitly does not cover higher molecular weight peptides and polypeptides, which is why peptide specification practice draws on the EMA Guideline on the Development and Manufacture of Synthetic Peptides (final, 2025-12-04) instead. That guideline expects batch-analysis summaries for all peptide batches in the dossier and cross-validation data where analytical methods are used, which is the same claim-to-readout chain the matrix rows test.
próximos pasos: what to request from a partner this quarter
Send the request list before the next technical call, not after it. Five items cover most of what the evidence matrix asks for.
-
Raw data retention policy. How long are chromatograms, mass spectra and integration records kept, and in what format can they be released?
-
Method validation summaries written against acceptance criteria that were predefined, not reverse-engineered to fit results.
-
System suitability records for the methods used on your program.
-
Pilot or engineering batch comparability data, so scale-up is shown rather than projected.
-
The batch-analysis summary covering all batches, including any that failed or were reprocessed.
If assembling that package is the bottleneck, MOL Changes supports analytical documentation from method development through release testing as one option among several capable suppliers. v. This article describes what a diligence-ready package contains, not what any specific regulator will require for a filing, and it is not legal or regulatory advice.
Frequently asked questions
But doesn’t a high purity number already prove the peptide is well controlled?
No. A purity percentage tells you how much of the sample eluted as the main peak; it does not tell you what the remainder is. That distinction matters because ICH Q3A(R2) sets impurity reporting, identification and qualification thresholds that scale with daily dose, so the same 97% figure carries a different obligation in a 2 mg dose than in a 20 mg one. Impurity control for peptides is a profile claim, not a percentage claim.
¿Temu̲ thogi nu'bu̲ ya di invertido ja 'nar socio 'mu̲i paquete ar delgado?
The gap is usually closable without restarting the program. Most missing artifacts turn out to be retention and documentation gaps rather than work that was never done, which means the fastest recovery path runs through the traceability layer and pilot comparability data. Ask for raw chromatograms, method parameters and batch records from work already completed before commissioning anything new.
ar guía ar péptidos sintéticos EMA da t'uni ja ma programa?
It is the modality-specific anchor for synthetic peptides in the EU. The EMA guideline on the development and manufacture of synthetic peptides expects batch-analysis summaries covering all peptide batches in the dossier, plus cross-validation data where analytical methods are used. Applicability depends on jurisdiction and filing route, and this is not regulatory advice.
Yogo'ä ICH Q6A hingi resuelve nuestras nt'a̲ni especificación?
Because ICH Q6A, published in December 2000, explicitly excludes higher molecular weight peptides and polypeptides from its scope. Peptide specification practice therefore draws on USP <1503> and the EMA peptide guideline instead, which is why a partner citing Q6A alone has not answered your specification question.
Njäts'i nu'bu
Record deal value is raising the bar for peptide development partner diligence because buyers now price manufacturing evidence, not molecule promise. The eight and a half billion dollars that went into peptide-therapeutics deals in 2024, described by PeptideStaff’s peptide therapeutics investment landscape review as the prior peak that 2025 activity was expected to surpass, set the terms of that shift: capital at this scale moves only against documentation a reviewer can test.
The broader change the article argues for is that diligence-ready evidence should be assembled continuously, not reconstructed under deal pressure. Reproducible synthesis records, a full impurity profile, orthogonal identity data, comparability runs and audit-ready traceability are cheaper to maintain as a byproduct of normal development than to rebuild in a data room on a deadline.
Treat the evidence matrix as a standing checklist rather than a one-time sprint, and request the analytical data package from your current or candidate peptide development partner this quarter.