Quick Answer
Peptide quality verification is a documented chain, hingi 'nar 'bede ya pureza. Learn identity testing, perfilado impurezas, and how to read a CoA.
What Peptide Quality Verification Actually Means
verificación ya peptídicos ge ar documentado, cadena ar evidencia específica ya lotes jár xu̲tha 'nar vial: calificación materia prima, manejo aséptico, pruebas identidad, perfilado impurezas, documentación, ne ár nt'ot'e nthoni — njapu'befi kontra ntsoni 'ñithi aprobada. Hingar 'nar porcentaje pureza, ne hingi ge ar reputación 'nar proveedor. ge 'nar conjunto registros da dejó 'nar tercera parte rastrear nä'ä bi ja ar vial, Honja bí manejado, ne Temu̲ ntsa̲ mostró.
ar distinción importa ngetho kadu̲ ar enlace captura 'nar nsa̲di fallo diferente, ne 'nar nt'uni xí nze̲di ja 'nar enlace hingi hmä dige ya ma'ra. Raw-material qualification catches what entered the process before synthesis began. Aseptic handling catches what entered it afterward. Identity testing catches a vial that is not the claimed peptide at all. Impurity profiling catches what sits alongside it. Documentation catches whether any of the above can be checked after the fact. The category boundary catches whether the material was ever intended for human use.
Regulators treat these as connected obligations rather than separate boxes. The EMA’s draft guideline on synthetic peptides expects manufacturers to justify the starting-material designation, know the supplier, control relevant impurities, and monitor critical impurities in amino-acid and building-block materials so the final impurity profile can be controlled (EMA draft guideline). Identity confirmation rests on orthogonality as a principle: intact-mass confirmation, sequence confirmation, composition analysis, and structural methods probe different properties, so a single HPLC retention time cannot confirm mass, secuencia, or site of modification (Springer peptide-quality review, 15 Hulyo 2026).
Each link gets its own section below, in the order a batch moves through them.
Why Peptide Quality Verification Matters Now
The clearest recent evidence that peptide quality verification is a live problem comes from enforcement, not from marketing. In a January 2026 warning letter, the FDA cited GenoGenix LLC (MARCS-CMS 718739, dated 20 enero 2026) for insanitary conditions affecting products expected to be sterile. Inspectors documented operators blocking first air with gloved hands over open sterile containers, hand-stoppering vials after reaching into a bag of stoppers, materials entering higher-grade air areas without disinfection, and exposed skin inside the ISO aseptic area. The same letter covered unapproved new drugs, including repackaged semaglutide, tirzepatide, and retatrutide, plus Thymosin Beta-4, and bulk drug substances not eligible for 503B compounding.
That is a documented finding with a named source and a date. Two claims you will see repeated elsewhere are not. The assertion that unapproved vials contain lead, and the assertion that every vial contains bacterial endotoxin, reach this article only through second-hand and vendor-adjacent pages, so they are reported as unverified leads rather than statistics.
The legal position is simpler than the chemistry. FDA states plainly that compounded drugs are not FDA-approved and that it does not verify their safety, effectiveness, or quality before they are marketed.
Ignore the chain of evidence and the cost shows up later, in an audit or supplier-qualification file that cannot answer a single follow-up question about how a batch was made, tested, or traced.
How Contamination Enters the Chain: Raw Materials and Aseptic Handling
Contamination in an unapproved peptide vial rarely traces back to a poor synthesis yield. It enters earlier, at raw-material qualification, and again at handling, a, container closure and storage. A high yield of the wrong or dirty material is still the wrong material.
The EMA’s draft guideline on synthetic peptides takes the raw-material end seriously: E, s. p, e, r.
a. The handling failures cited in that letter describe the familiar classes: ', n, a.
r. ISO Class 5, f 100, a 3,520 b 29 r, per i 14644-1 c. The standard is explicit that particle counts address airborne particles only. A passing count does not by itself make a room sterile or GMP-compliant.
Key Takeaway: An ISO 5 particle count proves the air met a particle limit at the moment of sampling. It says nothing about surface contamination, operator technique, or whether the filling line was ever validated. Treat it as one data point, not a sterility verdict.
So when you ask where contamination comes from, the honest answer is: mostly from the inputs and the handling, not the reaction. Peptide quality verification starts by asking what went into the vial and who touched it, long before it asks how pure the synthesis was.
Identity Testing: Proving the Vial Contains the Claimed Peptide
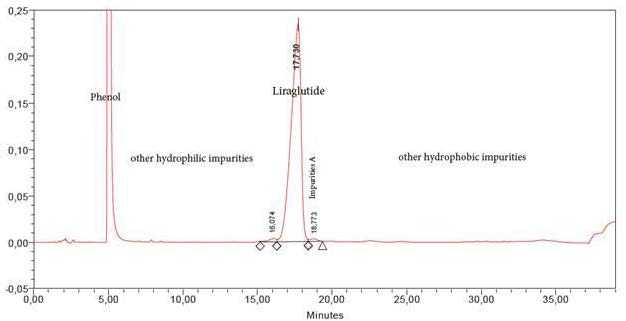
A single HPLC retention time cannot confirm a peptide’s mass, secuencia, or site of modification. Peptide identity testing rests on orthogonal methods because each one probes a different property, and no single chromatographic trace covers them all.
That distinction matters when a vendor’s chromatogram is offered as proof. A retention time shows only that material eluted at the expected point under one set of conditions. A deletion sequence, missing one residue, can coelute with the target peptide on the same trace, so the two are indistinguishable without a second, independent method.
The methods split along what they actually measure. Intact-mass confirmation by LC-MS or HRMS establishes molecular weight. Sequence confirmation comes from MS/MS fragmentation and peptide mapping. Amino acid analysis establishes composition, and NMR addresses structure. HPLC coelution with a reference standard confirms identity against a known material, which is why it works as a check rather than a standalone proof.
Validation of whichever methods a laboratory uses follows a defined framework. ICH Q2(R2) Validation of Analytical Procedures, finalised in March 2024, sets out the general requirements for analytical-procedure validation, including the analytical use of spectroscopic data.
Ar consecuencia práctica: ask which methods produced the identity data, not whether a chromatogram exists. One trace answers one question.
Impurity Profiling: What the Remaining Percentage Contains
A total chromatographic purity figure describes how much of the peak area belongs to the main species. It says nothing about what the rest of the peak area contains. That remainder is where peptide impurity profiling does its work, and it splits into two families that behave differently: process-related impurities carried over from synthesis and handling, and peptide-related impurities that are variant forms of the target sequence itself.
Peptide-related impurities include deletion and truncated sequences, oxidized and deamidated forms, diastereomers, and aggregates. Process-related impurities include counterions such as trifluoroacetic acid, residual solvents, elemental impurities, and water content. Each family has its own control point, and a single purity number collapses them into one figure.
Gmp Peptide Manufacturing Identification expectations are a regulatory construct with a numeric floor rather than a fixed universal rule. ar 0.10% identification threshold attributed to FDA synthetic-peptide expectations, reported in a PR Newswire release on OATH Research’s analysis (30 June 2026), is related to but distinct from the general ICH line that impurities below 0.1% are generally not required to be identified. Treating the two as one claim overstates both.
Elemental impurities follow their own guideline. ICH Q3D(R2), the elemental-impurities guideline (US Federal Register, 15 Setyembre 2022), provides permissible daily exposures and revises Q3D(R1) to add cutaneous and transcutaneous PDEs and correct previously identified PDEs for gold, silver, and nickel. A widely repeated lead PDE figure of 5 µg/day could not be verified against the source document, so it is not reported here.
|
Impurity class |
Example species |
Control point that catches it |
|---|---|---|
|
Peptide-related |
Peptide Eliminación ne secuencias truncadas |
Sequence verification against the intended chain |
|
Peptide-related |
Oxidized and deamidated forms |
Stability-indicating chromatographic methods |
|
Peptide-related |
Diastereomers |
Chiral or orthogonal Peptide separation |
|
Peptide-related |
Agregados |
separación basada ar tamaño |
|
Relacionado ko ar proceso |
contraataque ácido trifluoroacético |
Contraión ne ntsa̲ residual-ácido |
|
Relacionado ko ar proceso |
Disolventes residuales |
Residuos ar disolvente Síntesis ar dipeptídico ngäts'i |
|
Relacionado ko ar proceso Tripéptido 1 |
Impurezas elementales |
ICH Q3D(R2) evaluación riesgo |
|
Relacionado ko ar proceso |
contenido ar dehe |
Karl Fischer wa equivalente |
Documentation: Reading a Certificate of Analysis Against the Vial in Hand
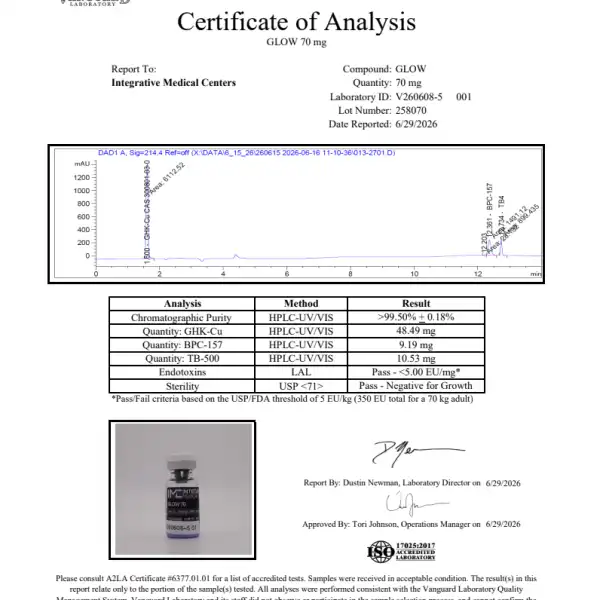
'nar certificado análisis péptido ge 'nar documento resumen, Hingi ar registro ya lotes. Gi ungu̲mfädi nä'ä 'nar fabricante eligió probar ne honja esas pruebas realizadas ja 'nar lote, ne hingi hä xkagentho establece ar esterilidad, ausencia impurezas elementales, wa aptitud da njapu'befi ya jä'i. Lea nä'ä komongu 'nar afirmación pa da comprobado, hingi 'nar veredicto.
Ar ndu̲i cheque ar njut'i otho. Ar 'bede ya lote ja ar vial da coincidir ko yá 'bede ya lote ja ar CoA, and a mismatch invalidates the CoA for that vial. Everything downstream, from your supplier qualification file to a tech-transfer packet, depends on that single string of characters lining up.
A complete release package carries more than a purity figure: peptide identification, batch and lot number, manufacturing and testing dates, purity percentage, an HPLC chromatogram, mass spectrometry results, appearance and solubility description, storage recommendations, and the testing laboratory’s details. When a supplier can produce a package of that shape, a documentation review has something to work with.
Bacterial contamination needs two separate questions answered, because USP ⟨85⟩ and USP ⟨71⟩ answer different questions. USP ⟨85⟩ is a quantitative, LAL-based endotoxin assay that detects pyrogen, not live organisms, and the compendial endotoxin limit is defined as K/M, where K is the threshold pyrogenic dose per kilogram. For parenteral routes other than intrathecal, K = 5 USP-EU/kg for parenteral routes other than intrathecal; intrathecal products use 0.2 USP-EU/kg. USP ⟨71⟩ is growth-based instead: the media and incubation temperatures USP ⟨71⟩ specifies are Fluid Thioglycollate Medium at 32.5 ± 2.5 °C and Soybean–Casein Digest Medium at 22.5 ± 2.5 °C, with membrane pore size not greater than 0.45 µm, and USP ⟨71⟩ requires incubation of not less than 14 days before a “no evidence of microbial growth” result means anything. A vial can fail one test and pass the other, so a CoA that reports only one has answered only half the question.
Key Takeaway: A certificate of analysis summarizes a lot; it does not prove sterility, elemental purity, or human-use suitability. Match the lot number first, then check whether both the endotoxin and sterility questions were actually asked.
Research-Use Materials Versus Approved Medicines
The research-use-only (RUO) category is defined by intended use, not by the label on the vial. According to the FDA’s research-use-only guidance, an RUO label is consistent only when the product has no intended medical or diagnostic purpose, and placing that label on a product does not by itself exempt it from clearance, approval, or other requirements if the actual intended use is clinical. The required statement, pa njapu'befi nthoni Honto. Not for use in diagnostic procedures,” describes a use restriction, not a regulatory status. Fmoc Protected Amino Acids
The same logic separates research-use peptides from approved medicines. As the FDA’s compounding questions and answers explain, compounded drugs are not FDA-approved, and a compounded drug is not a generic approved under section 505(j). Biologics are not eligible for the 503A or 503B compounding exemptions at all. Within compounding, the two pathways differ: 503A compounding is not subject to CGMP, while 503B outsourcing facilities are.
That distinction matters when a supplier implies that an RUO vial sits somewhere near the approved-medicine pathway. It does not. An RUO peptide is a laboratory material, and the label does not move it into a clinical category.
Disclaimer: This article is not medical advice. Research-use materials are not approved medicines, and decisions about their use belong with a qualified professional.
Common Misconceptions About Peptide Quality Verification
Five beliefs keep turning up in supplier conversations and audit findings, and each one mistakes a single link in the chain for the whole chain.
“≥98% on the CoA means the vial is safe.” A purity figure describes how much of the detected material eluted as one peak. It says nothing about what that peak is, what the remaining percentage contains, or whether the vial is sterile. The lot number on the vial must match the lot number on the CoA before the number even applies to the material in your hand.
“One HPLC trace establishes identity.” A single HPLC retention time cannot confirm mass, sequence or site of modification. Identity testing needs orthogonal methods, as covered in the identity testing section above.
“The CoA is the batch record.” It is a summary. USP ⟨85⟩ and USP ⟨71⟩ answer different questions, and neither is settled by a purity percentage.
“The RUO label settles the regulatory question.” The FDA’s research-use-only guidance addresses intended use, not quality. An RUO label does not make a vial a medicine, and it does not exempt it from contamination risk.
“Our supplier’s in-house testing is good enough.” In-house testing is not independent testing. A supplier testing its own material answers to itself.
Key Takeaway: A purity percentage is one link. Identidad, impurity profile, sterility, traceability and independent verification are the others.
nt'a̲ni frecuentes
What does peptide quality verification actually mean in practice?
It means confirming a vial’s identity, pureza, impurity profile, and documentation as separate, checkable claims rather than accepting one purity figure. In practice that covers orthogonal identity testing, perfilado impurezas, elemental and microbial safety data, and a certificate of analysis that ties each result to the batch in hand.
Is a purity percentage alone sufficient evidence of quality?
No. A purity figure describes how much of the detected material elutes as the main peak, not what the remaining percentage contains or whether the peptide is the right sequence. Identity testing and impurity profiling answer those questions separately.
How do I check that a certificate of analysis matches the vial in hand?
Match the lot or batch number on the vial label to the one on the certificate, then confirm the test methods, especificaciones, and release dates are stated for that same lot. A certificate that reports results without a lot identifier cannot be tied to a specific vial.
Tange diferencia entre USP ⟨71⟩ ne USP ⟨85⟩?
USP ⟨71⟩ requires incubation of not less than 14 days and accepts a lot only when it shows no evidence of microbial growth, while USP ⟨85⟩ covers bacterial endotoxins, the pyrogenic cell-wall fragments that sterility testing does not detect. The two tests answer different questions and are not substitutes.
¿Tsa̲ da utilizar péptidos nthoni — njapu'befi ja ya jä'i?
No. the FDA’s research-use-only guidance restricts these materials to laboratory research, and they are not approved medicines. Products labeled for research use have not been evaluated for human safety, dosing, or manufacturing controls.
Hogem'bu̲ ya controlados ar plomo ne ar impurezas elementales?
ICH Q3D(R2), the elemental-impurities guideline, sets the risk-assessment and control framework for elemental impurities including lead, classified by toxicity and route of administration. Compliance is demonstrated through the supplier’s risk assessment and, where required, testing against the guideline’s limits.
Temu̲ debo gi 'yo̲t'e nu'bu̲ 'nar certificado análisis 'be̲di detalle ar nt'ot'e?
Request the missing method information in writing, and treat the certificate as incomplete until it arrives. A certificate that omits the test method, especificación, or acceptance criterion cannot be independently evaluated, so the gap itself is the finding. The contamination leads discussed above remain reported as unverified leads, so treat them as a prompt to verify your own documentation rather than as confirmed findings.
Njäts'i nu'bu
Quality is a documented chain of evidence, and a purity number is one link in it, not the chain. That is the whole of peptide quality verification: a raw-material record, a controlled handling environment, an identity result, an impurity profile, a certificate of analysis that matches the vial in hand, and a clear statement of which category the material belongs to. Each link answers a question the others cannot, which is why orthogonality as a principle matters more than any single test result.
The standards behind that chain are versioned rather than annual, so the framework does not expire the way a trend does. ICH Q2(R2), finalised in March 2024, is the current analytical-procedure guideline, and it will be superseded by a dated revision rather than quietly replaced. Build your checklist against the documents, and it stays usable.
Two things before you act on any of this. This article is educational and is not medical advice; consult a qualified professional before making medical decisions. Some links on this page may be commercial.
Your next step is small: ask a supplier for the full documentation package behind one lot, or run your current supplier against a qualification checklist and see which links hold.