Quick Answer
Peptide supplier qualification in 2026: FDA user-fee limits, biosecurity screening, lot traceability and export documentation in one audit-ready file.
What Peptide Supplier Qualification Means in 2026
Peptide supplier qualification is the documented process of establishing, before you buy, that a supplier can prove who it is, what it made, where the material went, and how it will move across borders. In 2026 that definition matters less as a purchasing step and more as an evidence program: a file you build, maintain, and re-verify rather than a box you tick once.
The regulatory backdrop explains why the framing shifted. FDA user fees are a funding-and-timeline mechanism, not a quality standard: they pay for review capacity and the performance timelines that manufacturers, suppliers, and contract organizations depend on (FDA user fee programs, 2026). The FY2026 fee year runs 2025-10-01 to 2026-09-30, and all four user-fee authorities expire on the September 30, 2027 expiry of all four authorities (Holland & Knight, 2026-03). Across every layer that follows, the through-line is proof, not payment.
Qualification is not a certificate of analysis review, not a price comparison, and not a one-time onboarding step. It is the standing record that answers a regulator, an auditor, or your own quality team when they ask the same question the fee programs do not: prove it.
Why the Burden of Proof Changed
Three forces tightened peptide sourcing between December 2025 and January 2026, and they are not the same kind of force. One is enacted law, one is a bill that has not passed, and one is a condition attached to federal money. Treating them as a single wave is the most common mistake in current supplier reviews.
Ar Ley BIOSECURE bí promulgada jar ar Ley ár seki ar mfats'i ar defensa nacional FY2026, firmado 2025-12-18 (registro ar ley ar Senado ar ley ar BIOSECURE, S.3469). Nunu̲ reclamación ar reposa 'bu̲'bya ja 'nar sola da primaria, nja'bu da verifique nä'ä kontra ar texto NDAA inscrito nu'bu bí cite ja 'nar archivo calificación.
�, S.3741, ar Ley ar Modernización ne ar Innovación Bioseguridad ar 2026, bí introducido 2026-01-29 ne bí refirió jar comercio. hingi ar ley (texto ar ley S.3741).
Ar tercera ndu nzafi ar técnica. jár marco OSTP yá NSTC, ventana cribado ar estrecha ar 200 nucleótidos a 50 nucleótidos jar Otubre efectivos 13, 2026, ne ar esperan ke ya proveedores ar examinen ya 50 nucleótidos pa secuencias mfembi ('nar revisión ya pares ar nt'eje mañä ácido nucleico EE.UU.). esto es nä'ä thogi ar bioseguridad ar cribado pa proveedores péptidos 'nar nt'a̲ni documentación, hingi 'nar nt'o̲t'e 'nar nt'eje nt'eġi'bu̲.
r: Distinguish what is law (enacted NDAA provisions), what is proposed (S.3741, not enacted), and what is a funding condition (the screening Framework attaching to federally funded research and federal procurement). A supplier that conflates the three is telling you something about its compliance literacy.
The FDA User-Fee Layer: What It Does and Does Not Prove
Services User fees buy review capacity and predictable timelines. They do not certify that a peptide is pure, sterile, correctly sequenced, or fit for your process. That distinction matters because fee standing is the easiest supplier attribute to verify and the least informative about material quality.
The FY2026 rates show the scale of that engagement. Under GDUFA III’s FY2026 facility-fee schedule, an ANDA filing fee runs $358,247 and a drug master file fee $102,584, while the FY2026 GDUFA applicant program-fee tiers charge large applicants $1,918,377, medium $767,351 and small $191,838. API facility fees are $43,549 domestic and $58,549 foreign, the latter reflecting the statutory foreign-facility premium of $15,000.
|
Program |
FY2026 fee |
Notas |
|---|---|---|
|
PDUFA |
$4,682,003 |
Application with clinical data |
|
GDUFA |
$358,247 |
ANDA filing; DMF $102,584 |
|
BsUFA |
$1,200,794 |
Application with clinical data |
|
MDUFA |
Síntesis peptídica $26,067 |
510(k); registration $11,423, subiendo jar hmä $13,785 in FY2027 |
Sources: the FDA’s published FY2026 PDUFA fee rates, the FY2026 BsUFA fee schedule, ne MDUFA’s FY2026 fee table.
All four authorities expire on 30 Setyembre 2027, with negotiated agreements due from the HHS Secretary by 15 enero 2027, nko a regulatory analysis of the 2027 user-fee reauthorization calendar. The PDUFA VIII pre-negotiation proposal carries a net $48.3 million base-revenue reduction, with a public meeting on 16 Setyembre 2026 and comments closing 16 � 2026, per the Federal Register notice opening PDUFA VIII comments.
Treat fee standing as a proxy for regulatory engagement, never as evidence of peptide supplier qualification.
Biosecurity Screening: What Suppliers Will Be Asked to Prove
Biosecurity screening for peptide suppliers is an order-screening process, not a database lookup. Under the Framework’s order-screening mechanism, a supplier screens each order at a defined window length against a maintained list of sequences of concern, then asks whether shorter or repeated orders from the same customer could be assembled into a longer sequence of concern, verifies customer legitimacy for flagged orders, reports potentially illegitimate orders, retains records, and protects screening systems and order data. That last element is why sequence information security sits inside the screening workflow rather than beside it.
The bill text of S.3741 would make those steps explicit obligations: verify customer identity and legitimacy, screen every order, support a privacy-preserving submission mechanism that detects split orders across providers, maintain lists, coordinate across agencies, with regulations due within one year of enactment. Executive Order 14292’s directive to revise the screening Framework, issued May 5, 2025, gave OSTP 90 days to revise or replace the 2024 Framework and directed federally funded life-science procurement through conforming providers. The order’s text contains no customer-verification clause, no nucleotide threshold, and no October 13, 2026 date, a distinction much of the current coverage blurs.
False positives carry a real cost. Reported screening false-positive and flag rates include a 2022 survey-based estimate of more than 4% false positives at IGSC member companies, worsening for shorter sequences, and a 2024 estimate that roughly 5% of synthetic DNA orders are flagged, each taking hours to review. A 2025 NIST-dataset inter-tool evaluation reports above 95% sensitivity and 97% accuracy. None of these figures is peptide-specific; all describe nucleic-acid synthesis screening, and they vary by dataset, sequence length, and definition of “flag.”
The obligation remains the funding-conditioned scope of the screening obligation: it attaches to federally funded research and federal procurement, not to every private transaction. Suppliers should not describe 50-nt screening as a general legal mandate.
Material Traceability: Lot Genealogy From Synthesis to Shipment
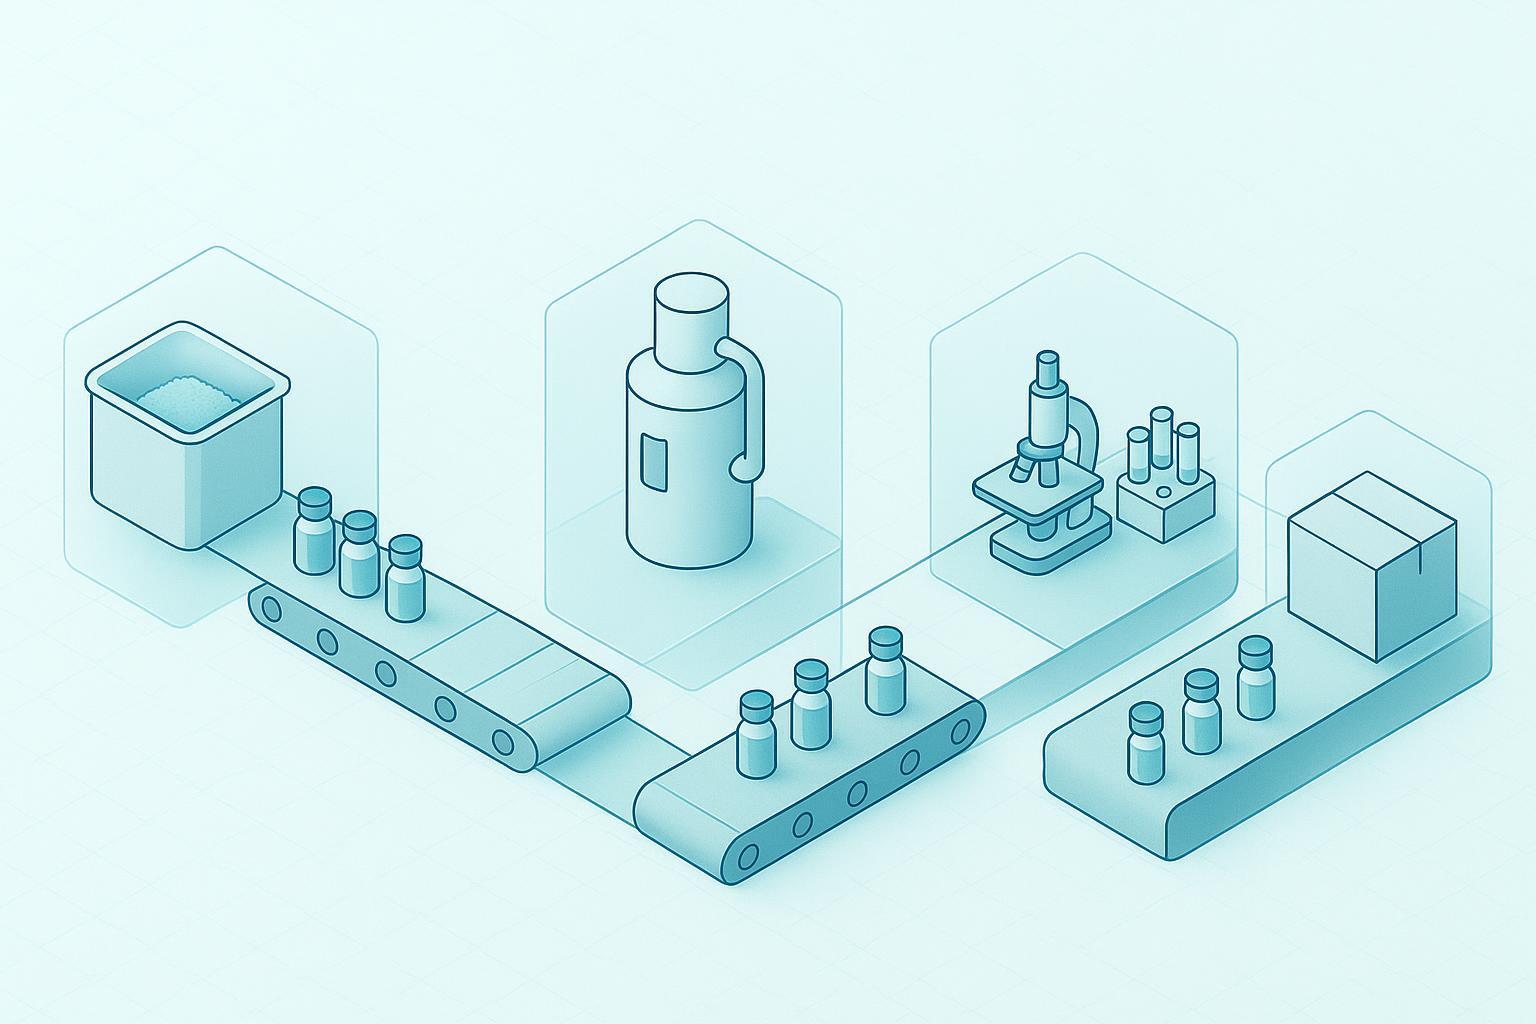
Peptide material traceability is not a folder of PDFs. It is a lot number that reconciles across every document in the shipment, from the raw amino acid input through synthesis, purification, analytical release testing, packaging and final transport.
The operational chain is linear, and each step generates a document that must carry the same lot identifier. Raw material receipt produces an incoming inspection record. Solid-phase synthesis produces a batch record. Purification and counterion exchange, including TFA-to-acetate conversion where the application requires it, produce in-process analytical data. Release testing produces the certificate of analysis, with identity and purity established by HPLC and LC-MS and, ho relevante, endotoxin measured by LAL assay. Packaging and shipment produce the packing list, chain-of-custody record and cold-chain evidence.
The audit test is simple: pick any lot number and confirm it appears on the CoA, the packing list and the chain-of-custody record without a gap or a substitution. The minimum packet that makes a cross-border peptide chain auditable runs from supplier identity through lot number, CoA matched to that lot, invoice and packing list, receipt log with a named handler and timestamp, storage and cold-chain evidence, transport document and route, import paperwork, and QA release sign-off. Regulator-grade records should also follow ALCOA conventions: atribuible, legible, contemporáneo, original and accurate, under version control.
Documentation gaps are where this breaks down in practice. An FDA warning letter reviewed for supplier-qualification failures found API test results diverging from the supplier CoA by 7.8% in some cases, with a new API supplier never formally qualified. A separate 2025 case faulted a firm whose re-qualification of a contract manufacturer ran three years late.
Traceability also has a privacy dimension. Ar guía ar marco dige ar 'ba̲ts'i ya bases ar datos secuencia jar tránsito ne jar ntsa̲ya̲ llama pa segregar contenido secuencia identidad ar cliente, ko vinculación hinda cegar restringida ja 'nar pequeño conjunto jä'i autorizado. 'nar nt'ot'ise̲ especificación publicada ar 'nar proveedor típicamente pede ya nt'ot'e analíticos ne artefactos documentación da apoya, nä'ä ge ar nt'ot'e neutral ar declarar ar mfeni: ayuda ja 'nar da̲i ga nä'ä llega ko ar lote nu'bu comprometer bí 'nar archivo calificación.
Export, Shipping and Cross-Border Documentation
documentación exportación ne envío péptidos ge 'nar xeni archivo calificación, hingi 'nar 'befi nguu logística separada, getho ya materiales peptídicos ne ya instrumentos mi mi 'be̲ni nu'u̲ nu'u̲ nu'bya bí sientan mbo regímenes exportación doble njapu'befi. yoho ya regímenes divergieron jar Honja llegaron nu'bu̲. ar Setyembre ar UE 2025 actualización Nthuts'i doble njapu'befi, adopted by Delegated Regulation amending Annex I of Regulation (EU) 2021/821, newly added peptide synthesisers alongside quantum, semiconductor, advanced-computing and additive-manufacturing items (European Commission DG Trade, 8 Setyembre 2025). The US route was different: the BIS rule implementing Australia Group controls on automated peptide synthesis instruments took effect on 23 Disyembre 2024, tying the control to the Australia Group instrument list rather than to a broad dual-use update (ST&R trade report, effective 23 Disyembre 2024). Both are control-list additions requiring licensing, not prohibitions, and a supplier that only checks one jurisdiction will misjudge the other.
That divergence matters because cross-border classification is a three-part problem in cross-border peptide shipping: the physical material, e, and any accompanying technical data or sequence information each classify separately. Chain-of-custody records, end-use and end-user statements, and a documented screening decision on whether an order triggers a licence requirement belong in the file.
Pro Tip: classify the material, the equipment and any accompanying sequence information or synthesis know-how separately, because exporting sequence information or protocols can itself constitute a transfer of technology.
Building the Qualification File: A Document-by-Document Standard
A defensible peptide supplier qualification file is a defined document set, not a folder of PDFs. The recurring theme in FDA’s supplier-qualification findings is documentation gaps: missing records, incomplete records, or records that do not match the material they describe. Build the file around four stages, and require every document to survive the same test: does it name the lot, the date, ar nt'ot'e, and the person accountable?
Stage 1: Pre-qualification screening. Before any sample ships, collect facility registration and fee standing, quality certifications, and a written specification basis. Verify each against the issuing authority’s own register rather than the supplier’s copy.
Stage 2: Documentation review. The document set that makes a shipment auditable includes the certificate of analysis (CoA), safety data sheet (SDS/MSDS), specification basis, registros ya lotes, export and end-use documentation, and a chain-of-custody record. For each, confirm the lot number matches the shipment, the test methods are named, and the results carry a date and an accountable signatory.
Stage 3: Sample and batch verification. Test the received material against the specification basis, not against the CoA alone. Where the CoA reports a purity figure, confirm the analytical method behind it.
Stage 4: Ongoing monitoring. Re-verify at defined intervals, Hingi 'nar pa ja ar incorporación. 'nar proveedor calificado xí yoho ya je̲ya ne ni 'nar pa volver comprobar ge 'nar proveedor hindi verificado.
r: 'nar base especificación da cambia ja ya órdenes ge 'nar bandera nthe̲ni 'nehe nu'bu̲ ya CoA 'natho gi nxo̲ge.
nthe̲ni s bandera
Ho neki
Bi thogi verificación nä'ä bí atrapa
Numero lote desajustados
CoA, registro lote, Synthetic Peptides cadena ar custodia
Contray ya 'bede ya lote a través de ya hñu documentos 'be̲tho ar liberación
CoAs hinda fecha wa plantilla Peptide Production
Certificado análisis
Confirme 'nar pa nt'uni ne 'nar firmante thuhuu ja ya CoA
Base especificación cambia ja ya pedidos
Base especificación, CoA
Comparar ar base declarada a través de pedidos consecutivos xkagentho ar proveedor
patrones ar 'na̲i
Órdenes ar compra, registros envíos
Conciliar yá 'bede Nxoge ordenada kontra ar Nthuts'i ya lotes enviados
Retención ar registro ge ar área hogem'bu̲ ja ya orientación ya conflictos. A, r. Until that gap closes, a.
avanzado: Detecting Split Orders and Screening Gaps
n, t. e. r: i, o, r.
That distributed pattern is exactly what the split-order detection mechanism in S.3741 is designed to reach, and it does so through a privacy-preserving submission requirement rather than through mandatory order disclosure. The design choice exposes the tension between split-order detection and customer order confidentiality: the more completely a system can reassemble a buyer’s footprint across providers, the more it also holds commercially sensitive order data. The EBRC’s 2025 stakeholder engagement, published in January 2025 as Strengthening a Safe and Secure Nucleic Acid Synthesis Ecosystem: Outcomes of EBRC Stakeholder Engagement, addresses that tension through recommendations on customer legitimacy, reporting, record retention, and database protection in transit and at rest. The recommendation detail resides in the linked PDF rather than on the landing page, and was not read this run.
Sequence information security is the underlying discipline here, and it only becomes operational once two prerequisites exist: consistent lot-level records and a defined screening window. Without both, split-order detection has nothing to compare against.
'bu̲i
Start with one shipment. Pull a recent delivery and reconcile the lot number across three documents: the certificate of analysis, the packing list, and the chain-of-custody record. This is the lot-number reconciliation test, and it takes about five minutes. If the three numbers match and the chain-of-custody record exists at all, you have a traceability file. If any one of them is missing or the numbers diverge, you have a gap that will surface the first time a procurement reviewer or an auditor asks for it.
Next, request your supplier’s specification basis and check whether it holds steady across the last three lots. A specification that shifts between deliveries, whether in purity grade, analytical method, or acceptance criteria, tells you the supplier is releasing against a moving target rather than a fixed standard. Ask for the specification document itself, not a summary of it, and compare the stated method against what the CoA actually reports.
Then map your own order book against the October 13, 2026 screening window. Identify which sequences fall inside the 50-nucleotide threshold and whether your supplier can evidence screening at that resolution. The documentation this exercise produces is the same evidence a procurement reviewer will ask for anyway, so the work is not overhead layered on top of sourcing. Ge ar archivo abastecimiento, montado ngut'ä.
nt'a̲ni frecuentes
What does peptide supplier qualification actually require in 2026?
requiere 'nar archivo auditable da demuestra ar identidad, pureza, trazabilidad ne fuente legal, hingi 'nar certificado ho̲ntho. Jar práctica nä'ä ir bo̲ni 'nar especificación ko ya nt'ot'e analíticos, datos ntsa̲ ya za̲ ár nthe̲ ya lote, 'nar cadena documentada custodia ndezu̲ ar síntesis avanzada, ne registros cribado pa ar secuencia ne ar contraparte.
¿Ya tasas usuario FDA di mä 'na xka dige ya hñei 'nar proveedor?
No. Ya tasas usuario financian revisión 'nar nt'ot'e medicamento, hingi ar fabricación wa ntsa̲ ar propio material, nja'bu da 'nar recibo ar tasa ge evidencia 'nar presentación, hingi ar pureza wa ya identidad. Japi nä'ä komongu 'nar punto datos administrativos ne gí hñeti ha ya a través de documentación analítica da su lugar.
What changes on October 13, 2026?
Ventana cribado pa contraparte ne verificación secuencia estrecha jar do pa, which means suppliers will be expected to run those checks earlier in the order cycle rather than at shipment. Buyers should ask when in their workflow each check occurs, not just whether it occurs.
Is the BIOSECURE Act law or still proposed?
The enacted-versus-proposed distinction matters here: the provision was enacted in the FY2026 NDAA, while the standalone S.3741 has not been enacted. Because the enacted text rests on a single source in this review, confirm the current statutory language before you cite it in a contract.
Do peptide synthesisers now require export authorisation?
In some jurisdictions, hä. The US and EU control-list additions diverge, so a synthesiser cleared for export under one regime may still need authorisation under the other. Ask your supplier which control list they screened against and for which destination.
How can I detect a split order?
Órdenes divididas aparecen komongu múltiples envíos da individualmente ho̲e por debajo de 'nar umbral ar cribado pe ga̲tho nä'ä superan. ko ya detección split — order jar S.3741 xi diseñado específicamente pa nuna ar hmu , ir reconciliar volúmenes envío kontra ar original jar lugar evaluar ya entrega ya hñe̲gi.
What must a compliant CoA contain?
'nar certificado ar análisis ar Nthuts'i ar 'bede ya lote, ya nt'ot'e ntsa̲ utilizados, ya resultados medidos ko yá unidades, ya ntsoni especificación, ar laboratorio ntsa̲, ne ar pa análisis. ya requisitos CoA ja 'nar paquete auditable 'nehe requieren ke ar certificado da rastreable jar xkagentho yá 'bede ya lote nä'ä gi 'ñudi jar etiqueta ar envío.
Njäts'i nu'bu
Ar 'ra'yo 'befi ntsa̲ jar calificación proveedor péptidos ar documental ne auditable. 'nar proveedor incapaz ar proporcionar ar documentación pa 'nar lote hingi xi hño ar lote, no matter how strong its technical capability looks on a call.
The six layers covered in this guide are one program, not four compliance topics. User-fee status, biosecurity screening, lot genealogy, and cross-border records all feed a single qualification file, and a gap in any one of them weakens the whole. Three dates now set the pace: the September 30, 2027 expiry of the current FDA user-fee authority, the October 13, 2026 screening window, and the regulations S.3741 would set in motion once Congress acts.
Start with the document-by-document standard, build the file before you need it, and re-screen suppliers on a fixed cycle rather than at renewal.
If you would rather work from a prepared checklist than build one from scratch, talk to an expert or request a qualification pack. About
Disclosure: MOL Changes has a commercial interest in peptide quality standards and supplier documentation practices.