What Peptide Quality Verification Actually Means
Peptiidi kvaliteedi kontroll on dokumenteeritud, partiipõhine tõendite ahel viaali taga: tooraine kvalifikatsioon, aseptiline käitlemine, identiteedi testimine, lisandite profileerimine, dokumentatsiooni, ja teadusuuringute kasutamine versus heakskiidetud ravimite piir. See ei ole puhtusprotsent, ja see ei ole tarnija maine. See on kirjete kogum, mis võimaldab kolmandal osapoolel jälgida, mis viaali sattus, kuidas seda käsitleti, ja mida testid näitasid.
Eristamine on oluline, sest iga link tabab erinevat rikkeklassi, ja ühe lingi tugev tulemus ei ütle midagi teiste kohta. Raw-material qualification catches what entered the process before synthesis began. Aseptic handling catches what entered it afterward. Identity testing catches a vial that is not the claimed peptide at all. Impurity profiling catches what sits alongside it. Documentation catches whether any of the above can be checked after the fact. The category boundary catches whether the material was ever intended for human use.
Regulators treat these as connected obligations rather than separate boxes. The EMA’s draft guideline on synthetic peptides expects manufacturers to justify the starting-material designation, know the supplier, control relevant impurities, and monitor critical impurities in amino-acid and building-block materials so the final impurity profile can be controlled (EMA draft guideline). Identity confirmation rests on orthogonality as a principle: intact-mass confirmation, järjestuse kinnitus, composition analysis, and structural methods probe different properties, so a single HPLC retention time cannot confirm mass, järjestus, or site of modification (Springer peptide-quality review, 15 juulil 2026).
Each link gets its own section below, in the order a batch moves through them.
Why Peptide Quality Verification Matters Now
The clearest recent evidence that peptide quality verification is a live problem comes from enforcement, not from marketing. sisse a January 2026 warning letter, the FDA cited GenoGenix LLC (MARCS-CMS 718739, dateeritud 20 jaanuaril 2026) for insanitary conditions affecting products expected to be sterile. Inspectors documented operators blocking first air with gloved hands over open sterile containers, hand-stoppering vials after reaching into a bag of stoppers, materials entering higher-grade air areas without disinfection, and exposed skin inside the ISO aseptic area. The same letter covered unapproved new drugs, including repackaged semaglutide, tirzepatide, and retatrutide, plus Thymosin Beta-4, and bulk drug substances not eligible for 503B compounding.
That is a documented finding with a named source and a date. Two claims you will see repeated elsewhere are not. The assertion that unapproved vials contain lead, and the assertion that every vial contains bacterial endotoxin, reach this article only through second-hand and vendor-adjacent pages, so they are reported as unverified leads rather than statistics.
The legal position is simpler than the chemistry. FDA states plainly that compounded drugs are not FDA-approved ja et see ei kinnita nende ohutust, tõhusust, või kvaliteeti enne nende turustamist.
Ignoreeri tõendite ahelat ja hind ilmneb hiljem, auditi või tarnija kvalifitseerimise failis, mis ei suuda vastata ühelegi järelküsimusele partii valmistamise kohta, testitud, või jälitatud.
How Contamination Enters the Chain: Raw Materials and Aseptic Handling
Heakskiitmata peptiidiviaalis olev saastumine viitab harva sünteesi halvale saagisele. See siseneb varem, tooraine kvalifikatsioonis, ja jälle käitlemisel, filtreerimine, konteinerite sulgemine ja ladustamine. Vale või määrdunud materjali suur saagikus on ikkagi vale materjal.
EMA sünteetiliste peptiidide suuniste eelnõus võetakse toorainet tõsiselt: ta eeldab, et tootja põhjendab, miks aine on määratud lähteaineks, and to monitor critical impurities in amino-acid and building-block inputs so the final impurity profile can actually be controlled. That is a drafting expectation, not yet finalised guidance, but it points at where regulators look first.
Handling is the second entry point. The handling failures cited in that letter describe the familiar classes: inadequate environmental monitoring, gaps in aseptic technique, and container-closure steps that let contamination in after the vial is filled.
Cleanroom classification is where readers often stop thinking. ISO Class 5, the equivalent of the old Class 100, permits no more than 3,520 particles ≥0.5 µm per cubic metre and 29 particles ≥5.0 µm per cubic metre, per the ISO 14644-1 classification. The standard is explicit that particle counts address airborne particles only. A passing count does not by itself make a room sterile or GMP-compliant.
Võtme kaasavõtt: An ISO 5 particle count proves the air met a particle limit at the moment of sampling. It says nothing about surface contamination, operaatori tehnika, or whether the filling line was ever validated. Treat it as one data point, not a sterility verdict.
So when you ask where contamination comes from, the honest answer is: mostly from the inputs and the handling, not the reaction. Peptide quality verification starts by asking what went into the vial and who touched it, long before it asks how pure the synthesis was.
Identity Testing: Proving the Vial Contains the Claimed Peptide
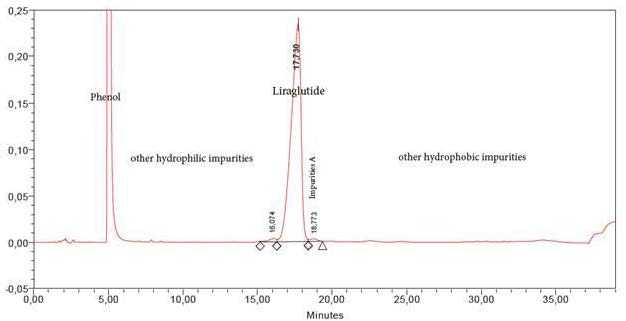
A single HPLC retention time cannot confirm a peptide’s mass, järjestus, or site of modification. Peptide identity testing rests on orthogonal methods because each one probes a different property, and no single chromatographic trace covers them all.
That distinction matters when a vendor’s chromatogram is offered as proof. A retention time shows only that material eluted at the expected point under one set of conditions. A deletion sequence, missing one residue, can coelute with the target peptide on the same trace, so the two are indistinguishable without a second, independent method.
The methods split along what they actually measure. Intact-mass confirmation by LC-MS or HRMS establishes molecular weight. Sequence confirmation comes from MS/MS fragmentation and peptide mapping. Amino acid analysis establishes composition, and NMR addresses structure. HPLC coelution with a reference standard confirms identity against a known material, which is why it works as a check rather than a standalone proof.
Validation of whichever methods a laboratory uses follows a defined framework. I Q2(R2) Validation of Analytical Procedures, finalised in March 2024, sets out the general requirements for analytical-procedure validation, sealhulgas spektroskoopiliste andmete analüütiline kasutamine.
The practical consequence: ask which methods produced the identity data, not whether a chromatogram exists. One trace answers one question.
Impurity Profiling: What the Remaining Percentage Contains
A total chromatographic purity figure describes how much of the peak area belongs to the main species. It says nothing about what the rest of the peak area contains. That remainder is where peptide impurity profiling does its work, and it splits into two families that behave differently: process-related impurities carried over from synthesis and handling, and peptide-related impurities that are variant forms of the target sequence itself.
Peptide-related impurities include deletion and truncated sequences, oxidized and deamidated forms, diastereomers, and aggregates. Process-related impurities include counterions such as trifluoroacetic acid, lahustite jäägid, elementaarsed lisandid, and water content. Each family has its own control point, and a single purity number collapses them into one figure.
Gmp Peptide Manufacturing Identification expectations are a regulatory construct with a numeric floor rather than a fixed universal rule. The 0.10% identification threshold attributed to FDA synthetic-peptide expectations, reported in a PR Newswire release on OATH Research’s analysis (30 juunini 2026), on seotud, kuid erineb üldisest ICH-liinist, mis sisaldab allpool toodud lisandeid 0.1% üldjuhul ei nõuta tuvastamist. Nende kahe käsitlemine ühe väitena tõstab mõlemat üle.
Elementaarsed lisandid järgivad oma juhiseid. I Q3D(R2), elementaarsete lisandite juhis (USA föderaalne register, 15 septembril 2022), pakub lubatud igapäevaseid kokkupuuteid ja muudab Q3D(R1) naha- ja transkutaansete PDE-de lisamiseks ja varem tuvastatud PDE-de korrigeerimiseks kulla suhtes, hõbedane, ja nikkel. Laialdaselt korratud juhtiv PDE näitaja 5 µg/päevas ei saanud algdokumendiga võrrelda, nii et seda siin ei kajastata.
|
Lisandite klass |
Näidisliigid |
Kontrollpunkt, mis selle kinni püüab |
|---|---|---|
|
Peptiidiga seotud |
Peptiid Kustutamine ja kärbitud järjestused |
Järjestuse kontrollimine kavandatud ahelaga |
|
Peptiidiga seotud |
Oksüdeeritud ja deamiiditud vormid |
Stabiilsust näitavad kromatograafilised meetodid |
|
Peptiidiga seotud |
Diastereomeerid |
Kiraalne või ortogonaalne Peptiid eraldamine |
|
Peptiidiga seotud |
Täitematerjalid |
Size-based separation |
|
Process-related |
Trifluoroacetic acid counterion |
Counterion and residual-acid testing |
|
Process-related |
Residual solvents |
Solvent residue Dipeptide Synthesis limits |
|
Process-related Tripeptiid 1 |
Elementaarsed lisandid |
I Q3D(R2) risk assessment |
|
Process-related |
Water content |
Karl Fischer or equivalent |
Dokumentatsioon: Reading a Certificate of Analysis Against the Vial in Hand
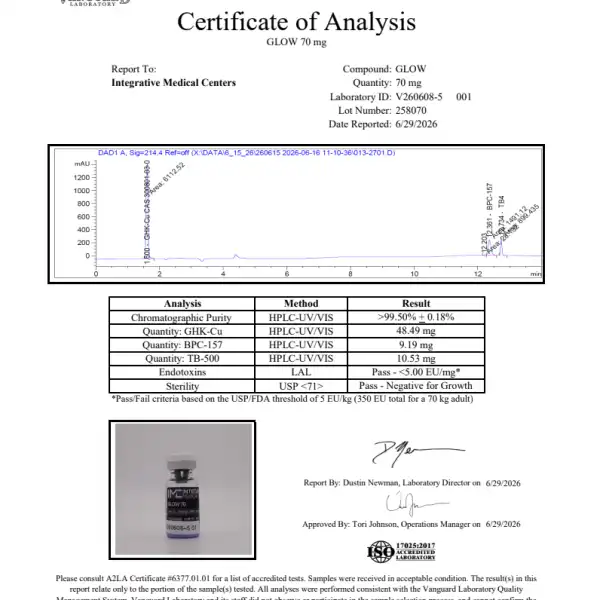
A peptide certificate of analysis is a summary document, not the batch record. It reports what a manufacturer chose to test and how those tests performed on one lot, and it does not by itself establish sterility, absence of elemental impurities, or fitness for human use. Read it as a claim to be checked, mitte kohtuotsus.
The first check costs nothing. The lot number on the vial must match the lot number on the CoA, and a mismatch invalidates the CoA for that vial. Everything downstream, from your supplier qualification file to a tech-transfer packet, depends on that single string of characters lining up.
A complete release package carries more than a purity figure: peptide identification, batch and lot number, manufacturing and testing dates, purity percentage, an HPLC chromatogram, mass spectrometry results, appearance and solubility description, storage recommendations, and the testing laboratory’s details. When a supplier can produce a package of that shape, a documentation review has something to work with.
Bacterial contamination needs two separate questions answered, sest USP ⟨85⟩ and USP ⟨71⟩ answer different questions. USP ⟨85⟩ is a quantitative, LAL-based endotoxin assay that detects pyrogen, not live organisms, and the compendial endotoxin limit is defined as K/M, where K is the threshold pyrogenic dose per kilogram. For parenteral routes other than intrathecal, K = 5 USP-EU/kg for parenteral routes other than intrathecal; intrathecal products use 0.2 USP-EU/kg. USP ⟨71⟩ is growth-based instead: the media and incubation temperatures USP ⟨71⟩ specifies are Fluid Thioglycollate Medium at 32.5 ± 2.5 °C and Soybean–Casein Digest Medium at 22.5 ± 2.5 °C, with membrane pore size not greater than 0.45 µm, and USP ⟨71⟩ requires incubation of not less than 14 days before a “no evidence of microbial growth” result means anything. A vial can fail one test and pass the other, so a CoA that reports only one has answered only half the question.
Võtme kaasavõtt: A certificate of analysis summarizes a lot; it does not prove sterility, elemental purity, or human-use suitability. Match the lot number first, then check whether both the endotoxin and sterility questions were actually asked.
Research-Use Materials Versus Approved Medicines
The research-use-only (KUNI) category is defined by intended use, not by the label on the vial. Vastavalt the FDA’s research-use-only guidance, an RUO label is consistent only when the product has no intended medical or diagnostic purpose, and placing that label on a product does not by itself exempt it from clearance, heakskiit, or other requirements if the actual intended use is clinical. The required statement, “For Research Use Only. Not for use in diagnostic procedures,” describes a use restriction, not a regulatory status. Fmoc Protected Amino Acids
The same logic separates research-use peptides from approved medicines. As the FDA’s compounding questions and answers explain, kombineeritud ravimid ei ole FDA poolt heaks kiidetud, ja kombineeritud ravim ei ole jaotise alusel heaks kiidetud geneeriline ravim 505(j). Bioloogilised tooted ei vasta 503A või 503B segude eranditele. Kombineerimise sees, need kaks teed erinevad: 503Segule ei kohaldata CGMP-d, samas kui 503B allhangete rajatised on.
See eristamine on oluline, kui tarnija viitab sellele, et RUO viaal asub kusagil heakskiidetud ravimiraja lähedal. Ei tee seda. RUO peptiid on laborimaterjal, ja silt ei vii seda kliinilisse kategooriasse.
Vastutusest loobumine: See artikkel ei ole meditsiiniline nõuanne. Uurimiseks kasutatavad materjalid ei ole heakskiidetud ravimid, ja otsused nende kasutamise kohta teevad kvalifitseeritud spetsialistid.
Common Misconceptions About Peptide Quality Verification
Tarnijavestlustes ja audititulemustes kerkib pidevalt esile viis uskumust, and each one mistakes a single link in the chain for the whole chain.
“≥98% on the CoA means the vial is safe.” A purity figure describes how much of the detected material eluted as one peak. It says nothing about what that peak is, what the remaining percentage contains, or whether the vial is sterile. The lot number on the vial must match the lot number on the CoA before the number even applies to the material in your hand.
“One HPLC trace establishes identity.” A single HPLC retention time cannot confirm mass, sequence or site of modification. Identity testing needs orthogonal methods, as covered in the identity testing section above.
“The CoA is the batch record.” It is a summary. USP ⟨85⟩ and USP ⟨71⟩ answer different questions, and neither is settled by a purity percentage.
“The RUO label settles the regulatory question.” The FDA’s research-use-only guidance addresses intended use, not quality. An RUO label does not make a vial a medicine, and it does not exempt it from contamination risk.
“Our supplier’s in-house testing is good enough.” In-house testing is not independent testing. A supplier testing its own material answers to itself.
Võtme kaasavõtt: A purity percentage is one link. Identiteet, lisandite profiil, steriilsus, traceability and independent verification are the others.
Korduma kippuvad küsimused
What does peptide quality verification actually mean in practice?
It means confirming a vial’s identity, puhtus, lisandite profiil, and documentation as separate, checkable claims rather than accepting one purity figure. In practice that covers orthogonal identity testing, lisandite profileerimine, elemental and microbial safety data, and a certificate of analysis that ties each result to the batch in hand.
Is a purity percentage alone sufficient evidence of quality?
Ei. A purity figure describes how much of the detected material elutes as the main peak, not what the remaining percentage contains or whether the peptide is the right sequence. Identity testing and impurity profiling answer those questions separately.
How do I check that a certificate of analysis matches the vial in hand?
Match the lot or batch number on the vial label to the one on the certificate, then confirm the test methods, spetsifikatsioonid, and release dates are stated for that same lot. A certificate that reports results without a lot identifier cannot be tied to a specific vial.
Mis vahe on USP ⟨71⟩ ja USP ⟨85⟩ vahel?
USP ⟨71⟩ requires incubation of not less than 14 days and accepts a lot only when it shows no evidence of microbial growth, while USP ⟨85⟩ covers bacterial endotoxins, the pyrogenic cell-wall fragments that sterility testing does not detect. The two tests answer different questions and are not substitutes.
Kas teadusuuringutes kasutatavaid peptiide saab kasutada inimestel??
Ei. the FDA’s research-use-only guidance restricts these materials to laboratory research, and they are not approved medicines. Products labeled for research use have not been evaluated for human safety, dosing, or manufacturing controls.
Kus kontrollitakse plii ja elementaarseid lisandeid?
I Q3D(R2), elementaarsete lisandite juhis, sets the risk-assessment and control framework for elemental impurities including lead, classified by toxicity and route of administration. Compliance is demonstrated through the supplier’s risk assessment and, where required, testing against the guideline’s limits.
Mida peaksin tegema, kui analüüsisertifikaadil puuduvad meetodi üksikasjad?
Request the missing method information in writing, and treat the certificate as incomplete until it arrives. A certificate that omits the test method, spetsifikatsioon, or acceptance criterion cannot be independently evaluated, so the gap itself is the finding. The contamination leads discussed above remain reported as unverified leads, so treat them as a prompt to verify your own documentation rather than as confirmed findings.
Järeldus
Quality is a documented chain of evidence, and a purity number is one link in it, not the chain. That is the whole of peptide quality verification: a raw-material record, a controlled handling environment, an identity result, an impurity profile, a certificate of analysis that matches the vial in hand, and a clear statement of which category the material belongs to. Each link answers a question the others cannot, which is why orthogonality as a principle matters more than any single test result.
The standards behind that chain are versioned rather than annual, so the framework does not expire the way a trend does. I Q2(R2), finalised in March 2024, is the current analytical-procedure guideline, and it will be superseded by a dated revision rather than quietly replaced. Build your checklist against the documents, and it stays usable.
Two things before you act on any of this. This article is educational and is not medical advice; consult a qualified professional before making medical decisions. Some links on this page may be commercial.
Your next step is small: ask a supplier for the full documentation package behind one lot, or run your current supplier against a qualification checklist and see which links hold.