How to Assess Transfer Readiness Before You Write the Protocol
USP <1224> txhais kev hloov txoj kev ua haujlwm raws li 'txheej txheem sau tseg uas ua rau lub chaw kuaj muaj cai' (chav txais khoom) siv txoj kev kuaj tshuaj ntsuam uas pib hauv lwm lub chaw kuaj (chav hloov chaw),xyuas kom chav ntawd 'muaj kev paub txheej txheem thiab peev xwm ua txoj kev tshuaj xyuas hloov chaw raws li xav tau' (USP-NF, <1224> Kev hloov pauv ntawm cov txheej txheem tshuaj xyuas). Nyeem qhov kev piav qhia ntawd raws li lus thiab lo lus nug npaj txhij teb nws tus kheej: koj tsis xa ib daim ntawv, koj tab tom txheeb xyuas ib qho chaw. So the inventory comes before the protocol.
Services The regulatory floor is set by ICH Q2(R2), which states that when a validated procedure moves to another laboratory, “a partial or full revalidation of the performance characteristics and/or comparative analysis of representative samples should be performed,” with justification required if no transfer experiments are run at all. FDA's 2015 analytical-procedures guidance defers transfer detail to USP <1224> rather than restating it, so the chapter, not the guidance, is your working text.
|
Item |
Cov pov thawj thov |
Peptide Synthesis Pass condition |
|---|---|---|
|
Instrument and detector |
Qualification records, wavelength or mass range |
Matches transferring unit’s configuration |
|
Column and lot history |
Lot numbers, age, Kev khaws cia |
Equivalent selectivity on a system suitability run |
|
Software and integration rules |
Version, audit trail, integration parameters |
Same rules applied to the same chromatograms |
|
Reference Khw standard |
Certificate, potency, expiry, kev khaws cia |
Traceable, in date, sufficient quantity |
|
Analyst experience |
Training records, prior peptide assays |
Documented competency on this technique |
Where any row fails, the verdict is one of three: transfer as written, transfer with pre-transfer remediation, or redevelop. That verdict is the core of peptide CDMO partner qualification, and it should be reached before a single protocol paragraph is drafted.
How to Write a Peptide Method Transfer Protocol That Holds Up
A method transfer protocol holds up when it names the transfer approach, the performance criteria, and the pass/fail rule before the receiving laboratory runs a single sample. USP <1224> recognises four approaches: comparative testing against predetermined acceptance criteria, co-validation between two laboratories, complete or partial validation, and a transfer waiver where the receiving unit has documented experience with a highly similar procedure. Which one applies depends on method criticality and complexity and on the receiving unit’s facilities and equipment.
ICH Q14’s analytical target profile sets the criteria before transfer begins. The ATP is the prospective summary of the performance a procedure must deliver, and comparative analysis of representative samples and reference materials is measured against it (ICH Q14, Step 4, 2023). Write the ATP into the protocol first, then derive thresholds from it.
Limitation: The replicate counts and thresholds below are worked examples calibrated to a stated risk level and intended use. Correct thresholds come from your own risk assessment. Confidence-level figures circulating in secondary sources could not be verified against a primary document, so this article states equivalence qualitatively.
Aligning Acceptance Criteria Across Two Peptide Laboratories
Acceptance criteria have to be agreed and signed before the first comparative run, not negotiated afterwards against whatever the data show. The defensible anchor is the analytical target profile: the receiving laboratory Synthetic Peptides should be able to demonstrate that its method meets the same performance requirement the sending laboratory’s method was validated against, rather than reproducing the sending laboratory’s numbers exactly.
One band is worth carrying into that discussion, with its limits stated. Regulated bioanalysis conventionally accepts accuracy within ±15% of nominal, widening to ±20% at the lower limit of quantification, with precision at or below 15% CV and 20% CV at the LLOQ (FDA, Bioanalytical Method Validation Guidance for Industry, May 2018). The ICH M10 text hosted by the FDA states the same precision limit (ICH M10, 2022). Treat this as a bioanalytical convention, not a universal peptide release criterion: the acceptance-criteria table itself sits in a PDF that could not be opened during research, so the worked thresholds above are the published convention rather than a verified extract of that table.
The practical point is that one threshold cannot govern five method types. Release, in-process, kev ruaj ntseg, cleaning and identity methods answer different questions, and a single borrowed number applied across all of them is the most common way analytical method transfer acceptance criteria become indefensible at audit.
Peptide-Specific Failure Modes That Generic Transfer Guidance Misses
Generic peptide method transfer guidance assumes the molecule survives the move intact. Four mechanisms specific to synthetic peptides break that assumption.
Resin and coupling chemistry differences shift the impurity profile. Two sites running the same sequence on different resins, or with different coupling reagents and activation conditions, produce different deletion and truncation populations. A gradient optimized for one impurity cluster will not resolve the other, so the receiving lab’s chromatogram looks worse even when the method performs correctly.
TFA counterion exchange distorts mass balance and purity readout. As Erckes et al. note in Towards a Consensus for the Analysis and Exchange of TFA Counterions (Pharmaceuticals 18(8):1163, published August 2025), solid-phase synthesis depends on trifluoroacetic acid as both cleavage agent and ion-pairing reagent, so peptides are obtained as TFA salts, and the counterion introduces a weighting error through salt-form molecular-weight differences. The same lot can therefore report different counterion-related values at two labs.
Disulfide scrambling changes identity and purity simultaneously. For cysteine-containing peptides, a shift in pH, oxygen exposure or dwell time during transfer can redistribute disulfide connectivity. The main peak may persist while the impurity profile and the identity assay diverge.
Co-eluting hydrophobic impurities need an explicit integration rule. Without a written rule, two analysts integrate the same unresolved hump differently.
The measurable part is the assay itself. Erckes et al. report quantification limits of 1.52 µg/mL for HPLC-ELSD, 20.68 µg/mL for 19F-NMR and 713 µg/mL for FT-IR, and note that prior published methods often lack validation according to ICH guidelines and often report TFA− content without assessing the exchanged ion.
Qhov tseem ceeb: Counterion quantification limits differ by more than two orders of magnitude between techniques, so a counterion discrepancy between two labs may be a technique mismatch rather than a manufacturing defect.
The Documentation and Tacit-Knowledge Layer Sponsors Underestimate
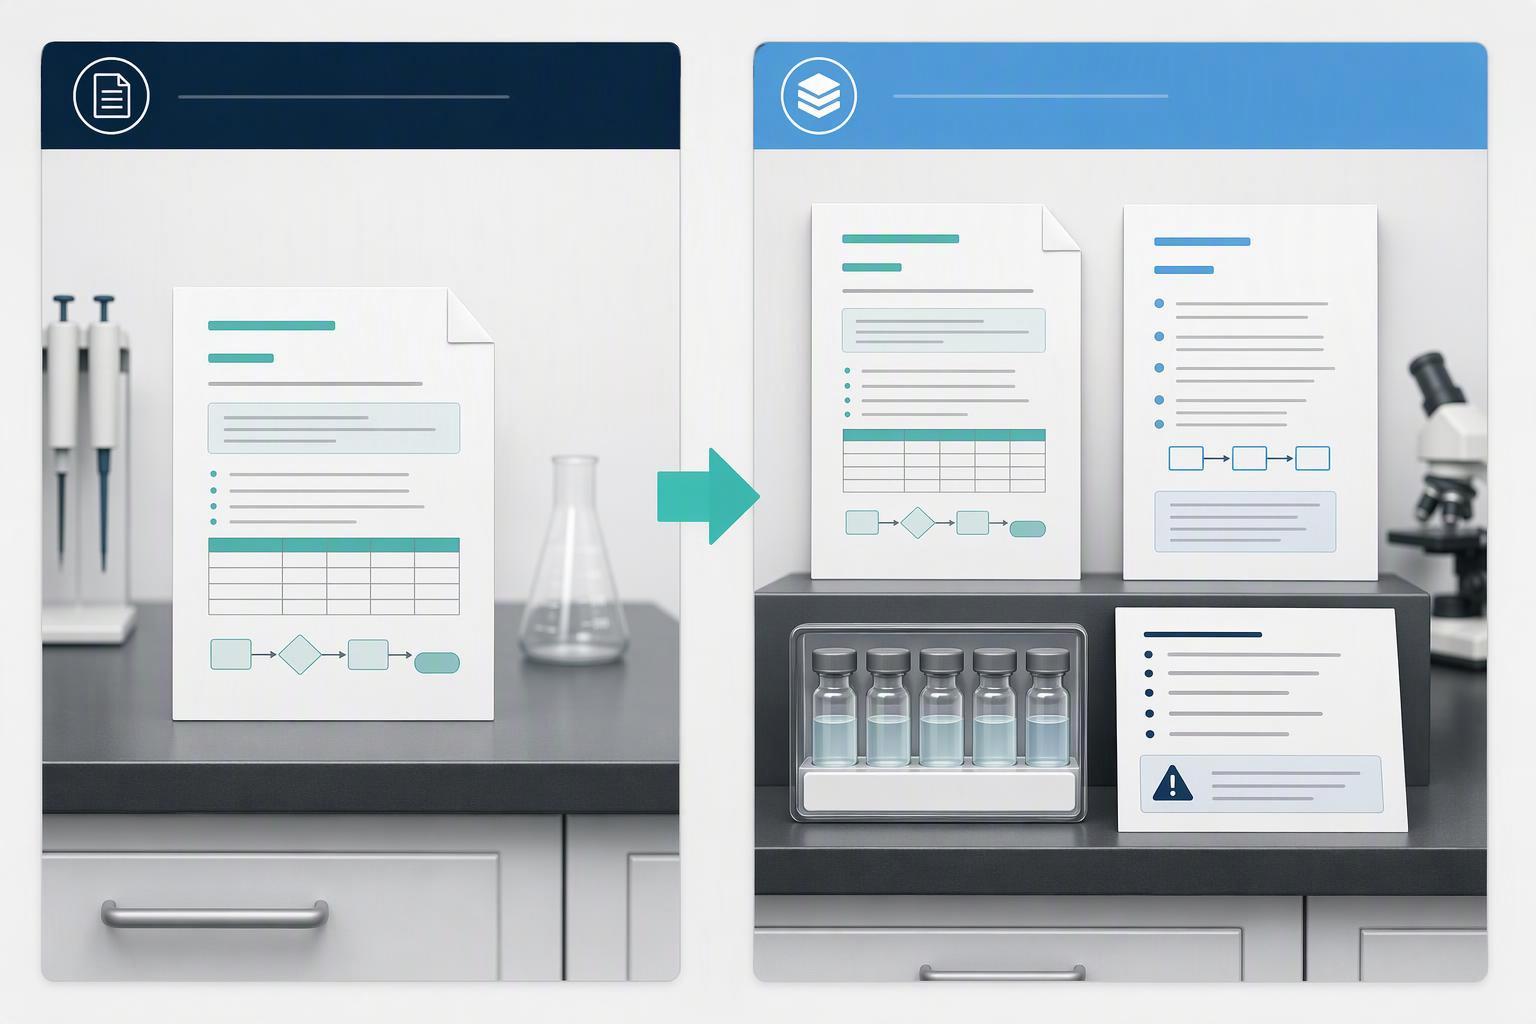
A method package can pass every completeness check and still fail in the receiving lab, because the artifacts that make a method transferable are rarely the ones written down. In peptide technology transfer, the gap is usually not the protocol itself but the reasoning behind it: why the integration rule for the hydrophobic impurity cluster was set where it was, which column lot behaved differently, which gradient adjustment was made and never recorded.
Three deliverables close most of that gap. Demand them before the protocol is signed:
Demand list: three artifacts a transfer-ready package must contain
A documented integration-rule set for the hydrophobic impurity cluster, including the decision logic, not just the final threshold About
A reference-standard qualification record covering both the sending and receiving site
A written list of known failure modes, each paired with the sequence context that triggers it
The pattern behind these failures is well documented in biopharmaceutical manufacturing generally. BioPlan Associates’ Fifth Annual Report and Survey of Biopharmaceutical Manufacturing Capacity and Production found an average of one batch failure every 40.6 weeks, with nearly 60% of respondents reporting their most recent failure within the previous 3 to 12 months (BioProcess International, 2008 survey). That figure covers manufacturing broadly rather than transfer specifically, so treat it as context for how often process knowledge gaps surface, not as a transfer failure rate.
The deeper constraint is talent. Experienced analytical staff hold much of this knowledge tacitly, and a package that never captures it transfers only the paperwork. Linking named personnel from the sending site to the receiving site for the duration of the comparison is the most reliable way to prevent that knowledge gap from opening.
How to Investigate an Out-of-Trend Comparative Result
Treat an out-of-trend comparative result as an analytical problem before you treat it as a manufacturing problem. The divergence itself does not tell you which it is; the investigation does. Branch on where the difference sits: the method, the sample, the standard, or the analyst.
|
Observation |
Likely cause class |
Immediate action |
Stop condition |
|---|---|---|---|
|
Single replicate outside the acceptance interval, other replicates pass |
Analyst or instrument variability Peptide Production |
Repeat the injection sequence with a fresh preparation |
Repeat passes within the interval |
|
Systematic bias across all replicates, both labs |
Method or standard |
Widen the comparative set; check standard qualification and equivalence |
Bias persists after standard verification |
|
Divergence only in the impurity profile |
Sample handling or integration rules |
Re-examine integration parameters and sample prep |
Profile matches after rule alignment |
|
Divergence survives repeat, widened set, and rule alignment |
Method itself |
Invoke co-validation |
Co-validation fails; remediate the method |
The escalation options here are the ones the approach taxonomy names: repeat, widen the comparative set, co-validation, or remediation. Skipping transfer experiments is not among them, and it is not defensible. The justification requirement means you must be able to show why each experiment was performed or omitted, and “we did not run it” is not a justification.
What Success Looks Like and What to Do Next
A peptide method transfer is complete when the comparative results fall inside the criteria you set before the first sample was run, at both sites, and the receiving unit can be described as qualified in the sense USP <1224> gives that word. Three artifacts should exist alongside those results: a signed reference-standard record held by both laboratories, a written list of the known failure modes for this molecule handed to the receiving team, and the comparative data set itself, traceable to the protocol version that produced it.
If any of those is missing, the transfer is not finished, whatever the results show. A passing data set with no signed standard record leaves the receiving lab unable to defend its next run.
The next step is a conversation about the specific gaps you have identified, not a general capability pitch. Bring your comparative data set, your criteria table and your failure-mode list to a discussion with an analytical team that works on peptide transfers routinely, thiab sib piv seb lawv yuav tswj hwm li cas rau ob peb qhov chaw uas koj cov txiaj ntsig ze rau qhov txwv. Qhov piv no feem ntau pab tau ntau dua kev tshuaj xyuas dav dav ntawm cov neeg muag khoom.
Disclosure: MOL Changes ua haujlwm hauv cov qauv zoo peptide thiab kev tshuaj xyuas, yog li peb muaj kev txaus siab lag luam hauv kev nkag siab txog qhov teeb meem no. Tsab ntawv no piav txog kev tshuaj xyuas thiab kev tswj hwm, tsis yog kev qhia kho mob. Kev txiav txim siab txog kev xa ntawv yuav tsum tau lees paub nrog lub koom haum kev noj qab haus huv thiab ib tug kws tshaj lij tswj hwm.
Cov Lus Nug Nquag Nug
Can a peptide method transfer happen without comparative testing?
Tsis yog ua ib qho kev sib piv uas muaj pov thawj tiv thaiv. ICH Q2(R2) txuas qhov tseeb ntawm txoj kev ua haujlwm nrog cov pov thawj sau tseg tias nws ua raws li lub hom phiaj hauv lub koom haum txais, thiab kev sim piv txwv yog txoj kev tsim cov pov thawj ntawd. T, u. s, a transfer report with no comparative data is an assertion, n.
How many replicates are enough?
e, e. g, not from a default number copied out of a template. t. x.
Yuav ua li cas yog tias pawg kab ntawm lub chaw kuaj txais hloov chaw khaws cia ntawm pawg impurity hydrophobic?
Saib nws raws li ib qho lus nug txog kev ruaj khov ntawm txoj kev ua ntej koj saib nws ua ib qho kev ua tsis tiav. Tsim kom paub tias qhov hloov pauv puas nyob hauv qhov kev xaiv uas txoj kev tau lees paub kom haum. Yog tias nws yog, sau tseg qhov hloov thiab qhov daws tau ua tiav, thiab lees paub tias pawg impurity tseem koom ua ke raws li ib pawg tau teev tseg. Yog tias nws tsis yog, chav kuaj txais tau cov tshuaj lossis pawg tsis sib npaug rau lub chaw kuaj hloov, thiab qhov kho yog nyob hauv txoj kev lossis kab ntawv qhia, tsis yog hauv cov qauv lees txais.
Puav leej puas muaj peev xwm tiv thaiv ib qho kev tso cai hloov chaw rau ib txoj kev ua haujlwm zoo sib xws heev?
Qee zaus, thiab kev tiv thaiv nyob ntawm qhov laj thawj, tsis yog qhov zoo sib xws. Kev tso cai yog muaj zog tshaj thaum txoj kev, cov cuab yeej, kab tshuaj lom neeg, thiab kev txawj ntse ntawm tus kws tshuaj ntsuam pom tau tias zoo ib yam, and when the sponsor documents why comparative testing adds nothing. Similarity alone is not a justification. Sponsors weighing this against a full comparative transfer often fold it into peptide CDMO partner qualification discussions, because the receiving lab’s own qualification records are what make the waiver arguable.
Yuav tsum ua li cas thiaj tswj tau qhov txawv ntawm cov txheej txheem nyeem tawm ntawm cov txheej txheem?
Compare like with like, and say which technique produced each number. Different analytical techniques can report a peptide’s counterion content on different bases, so a direct numerical comparison across techniques can show a difference that is an artifact of the readout rather than a real change in the material. In a comparability argument, either use the same technique on both sides or state the technique alongside each result and explain the basis of the comparison.
Xaus lus
Running a peptide method transfer without breaking the project comes down to sequence. Assess readiness before writing anything, so the protocol is built on what the receiving lab can actually execute. Write acceptance criteria both laboratories have signed, with the replicate counts and thresholds stated as numbers. Treat the peptide-specific failure modes, resin and coupling chemistry differences, counterion exchange, disulfide scrambling, hydrophobic impurity integration, as the load-bearing content rather than an appendix. Then close the documentation and tacit-knowledge gap, because that is where transfers quietly break: the method is sound, the paperwork is complete, and the knowledge that made the original method work never left the sending bench.
The single next action is the readiness assessment. Khiav nws ua ntej daim ntawv cog lus muaj, tsis yog tom qab thawj zaug sib piv tsis ua tiav. Yog tias daim ntawv teev qhov tsis sib haum rov qab ntev dua li lub sijhawm tso cai, yog lub sijhawm los tham nrog pab pawg tshuaj xyuas txog yam uas hloov tau raws li sau tseg thiab yam twg yuav tsum kho ua ntej hloov.