Тиз җавап
What a minimum connected data package for a peptide project must contain: чисталык, үзенчәлек, эчтәлеге, stability and assay records that cross-reference.
What “minimum connected data package for a peptide project” means in practice
Пептид синтезы The definition: the minimum connected data package for a peptide project is the smallest set of cross-referenced records that lets a second lab reproduce the result and a reviewer defend the decision.
Connected is not the same as complete. A complete package holds every document; a connected one makes each document point at the others, so a lot record names its chromatogram, the chromatogram names its method, and the method names the standard it was run against. MOL Changes draws the reproducibility test at re-runnability: if the package answers your questions but cannot be re-run by a second lab from the written record alone, ул процесс түгел, ә нәтиҗәне документлаштыра. That is the failure mode this guide is built to prevent.
What connected records look like in practice is set out in a worked example of a complete evidence package: synthesis route with impurity rationale, бер үк лотта чыгару һәм тотрыклылык турында мәгълүмат, a method-transfer report carrying acceptance criteria alongside actual results, and a certificate whose raw data remain retrievable.
Five claims sit inside that structure and must never substitute for one another: чисталык, үзенчәлек, эчтәлеге, тотрыклылык, һәм аналитик яраклашу. The rest of this guide takes them one at a time.
The five claims a package must keep separate
Чисталык, үзенчәлек, эчтәлеге, stability and assay suitability are five different claims, and each needs its own evidence. Conflating them is the most common review failure, and it usually starts with reading one purity number as a full characterization.
A published methodological recommendation for peptide handling sets the convention: a purified research peptide is typically specified at greater than 95% chemical purity, supported by preparative RP-HPLC purification and three distinct QC steps, analytical RP-HPLC for percent purity, MALDI/ESI-MS or MS/MS for mass and sequence confirmation, and amino acid analysis for concentration and composition accuracy (Recommendations for the generation, санлаштыру, storage and handling of peptides). The separation is not a house preference. I Q2(R2), июнь аеннан 2024, names assay/potency, чисталык, impurities and identity as distinct analytical purposes, һәрберсенең үз тикшерү өметләре бар, һәм USP peptide reference standards and impurity standards treat process-related and degradant impurities as separate categories rather than one purity figure.
The distinction between peptide purity vs identity is the one reviewers test first, because the two are measured by different instruments and answer different questions.
|
Кл Синтетик пептидлар aim |
Evidence requirement |
What it does not prove |
|---|---|---|
|
Чисталык |
Analytical RP-HPLC area percent |
Cannot distinguish co-eluting truncation from oxidation without orthogonal evidence |
|
Шәхес |
Mass confirmation against theoretical mass |
Does not establish purity or content |
|
Эчтәлек / чиста пептид |
Аминокислота анализы, counterion and water accounted for |
Area percent is not a content value |
|
Тотрыклылык |
Timepoint data under stated conditions |
Says nothing about identity or content at release |
|
Уңайлыкны тикшерегез |
Method-specific qualification |
Does not transfer to a different assay format |
The limitation matters more than the table. A purity area percent is not a content or net-peptide value, because counterion, water and residual solvent mass are not accounted for in the chromatogram. A lot can read 97% pure and still deliver materially less peptide than the label implies.
Sequence records and modification history
The sequence record is the package’s primary key. Every other record (хроматограмма, масса спектры, тотрыклылык протоколы, assay result) is only interpretable against a specific version of it, so it has to be version-controlled before any of those records mean anything.
A reviewer needs six fields to use it: эзлеклелек версиясе, N.- and C-terminus annotation, модификация сайты һәм химия, каршы форма, and lot ID. The modification field carries the most weight, because a modified residue is where a package most often loses traceability. A due-diligence playbook for peptide CRO selection treats the prior batch record or CoA for a sequence carrying the same modification, plus the analytical method used to confirm modification position, as part of the documentation demand list, not an optional extra.
Where a modification has no ready-made analytical route, reference standards do the work. USP supplies peptide API Reference Standards, related impurity Reference Standards and Analytical Reference Materials, plus system-suitability standards such as the Oxytocin Identification Reference Standard and Goserelin System Suitability Mixture, usable to develop and validate methods and to control drug substance and product batches (USP Biologics, Peptide Standards). Пептид җитештерү
The failure mode is a sequence edit with no immutable version trail. It is the most common cause of a non-reproducible experiment: the lot is real, the data is real, and nobody can prove which molecule was made.
Purity and identity records: what the chromatogram and mass spectrum actually prove
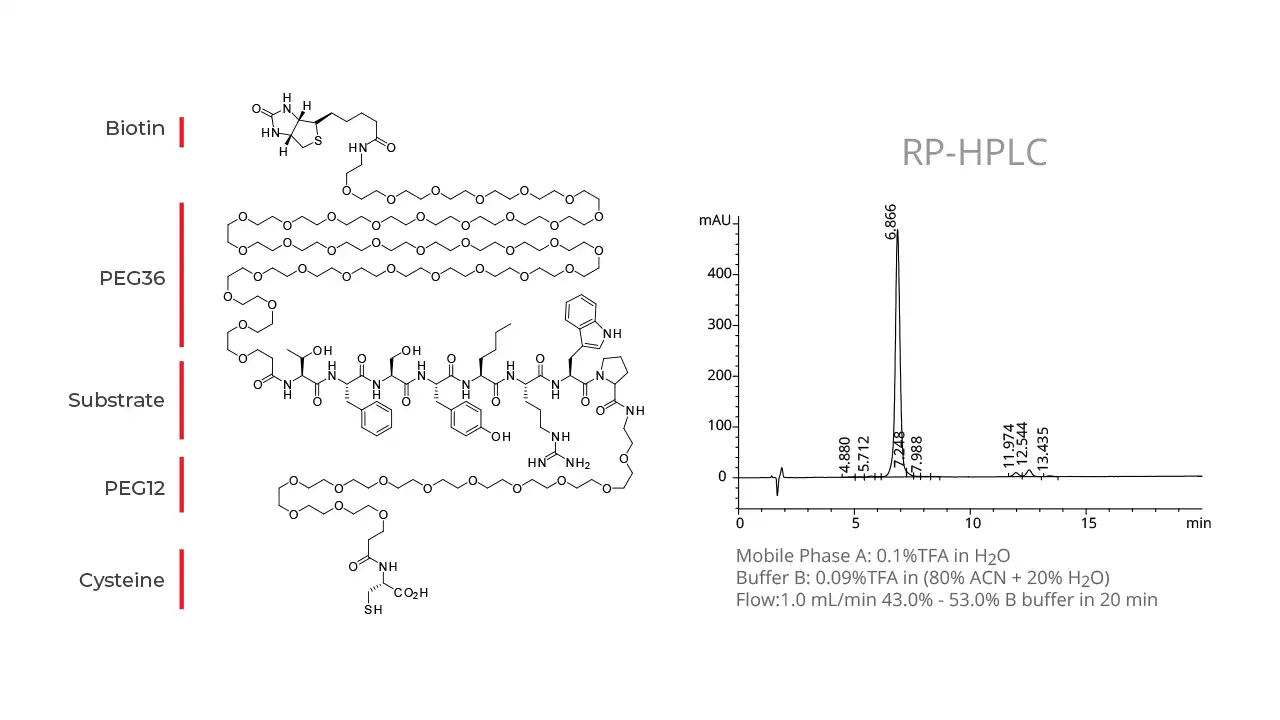
RP-HPLC establishes relative purity; ESI-MS establishes identity. Neither one establishes the other, and a package that treats them as interchangeable leaves the reviewer unable to answer either question.
The distinction is not editorial. I Q2(R2), июнь аеннан 2024, replaced Q2(R1) and treats assay/potency, чисталык, impurities and identity as separate analytical purposes, each with its own validation expectation. ICH Q2(R2) validation guidance as issued by FDA мартта 2024 carries the same structure. So the reviewer’s first question is not “is there an HPLC trace” but “which purpose does this method serve, and was it validated for that purpose.”
For the chromatogram, the fields that matter are column and dimensions, градиент, дулкын озынлыгын ачыклау, белешмә стандарты, and method ID. A leading shoulder at 0.6 RRT is only interpretable against those parameters: without the gradient, the retention time means nothing.
An analysis of peptide CRO sourcing practice states that buyers should expect HPLC and mass-spec data as standard rather than as an upcharge, and should be able to request raw QC data rather than a summary.
Two limits belong here. A summary table is not the chromatogram. And a mass spectrum confirms mass, not sequence position: a deletion and a substitution can return the same nominal mass, so MS alone cannot tell you where a sequence differs.
Эчтәлек, counterion and mass balance
Эчтәлек, яки чиста пептид, is a mass-balance value: it states how much of the weighed powder is actually peptide, after counterion, water and residual solvent are subtracted. It is the claim most often missing from a package that otherwise looks complete, and an HPLC area percent cannot stand in for it. Area percent measures the share of UV-absorbing peak area at one wavelength; it says nothing about the mass of material that peak represents.
A reviewer reading a purity-only certificate does the arithmetic anyway. If a lot is reported at 95% purity by area and the net peptide content is 70%, then a 10 mg weigh-out delivers roughly 7 мг пептид, and every downstream molar calculation inherits that gap. Counterion is usually the largest single term: acetate and trifluoroacetate both add mass that is not peptide.
Amino acid analysis is the reference approach for concentration and composition accuracy, because it quantifies peptide mass directly rather than inferring it from a chromatographic response (Journal of Pharmaceutical Analysis, 2016).
Ачкыч юл: Purity by area percent and content by mass balance answer different questions. A package that reports only the first leaves the reviewer to guess the second.
Where a package reports content but not the method behind it, the number is not verifiable, and the section of the certificate that should carry it stays empty.
Stability records: дизайн, conditions and timepoints
A stability result table is only interpretable when the protocol that produced it travels in the same package. Without the protocol, a receiving lab cannot tell whether the study applies to its own storage and handling conditions.
The required contents of a stability protocol are set out in Health Canada’s guidance on stability, sections 52 to 62 (2025-09-04). A peptide stability data package should carry, кулланылганда: product identification and master-document references; container and closure system type and size, plus total container count; lot or batch code with manufacture date and proposed expiry; sample size and test intervals per parameter; storage conditions with tolerances and how those tolerances are maintained; sample orientation reflecting worst case; specifications with acceptance criteria per parameter; the method of data evaluation, including statistical analysis or trending; and the report format.
These fields sit inside a framework rather than replacing it. .Әр сүзнең current Q1A(R2) stability framework dates to November 2003, һәм the Q5C expectations for peptide-like products (Июль 1996) state that products whose active components are typically proteins or polypeptides are particularly sensitive to temperature, оксидлашу, яктылык, ionic content and shear, so stringent storage conditions are usually necessary. Q5C does not supersede Q1A(R2); the two apply together. The field list above is a general stability-protocol requirement, not a peptide-specific or FDA-specific one.
Assay suitability and the records that make a result transferable
Assay suitability belongs to the method in the matrix, not to the peptide. A lot that reads clean by RP-HPLC can still fail a cell-based potency assay, because the assay measures what the peptide does in a buffer, a cell line and a plate, and those conditions are part of the result. The records that make a potency number transferable are therefore the format, the matrix, the controls and the acceptance criteria, not just the value.
For conjugate-type programs the bar is higher. ChemExpress sets out the analytical demands for peptide drug conjugate programs as LC-MS/MS for structural confirmation, SEC-MALS for aggregation, HPLC-UV/MS for drug-to-peptide ratio and free payload, peptide mapping for modification sites, and bioassays for potency, with methods validated per ICH guidelines. The same source states that without conjugation-site, drug-to-peptide ratio and aggregation characterization, “you cannot file an IND.”
Where a discovery-stage package may stop short: if the program is not yet filing, a documented method with matrix and controls can be enough, provided the record says which of those characterizations are still open.
Язмалар, audit trails and what makes a package defensible
A package is defensible when its electronic records meet the same integrity expectations as paper ones, һәм өлеш 11 states those expectations directly. 21 CFR 11.10(д) and the audit-trail requirement куркынычсызлыкка чакыра, компьютерда ясалган, Оператор язуларының датасын һәм вакытын мөстәкыйль язып торган вакыт маркалы аудит юллары, үзгәртү, or deleting electronic records, with changes never obscuring previously recorded information and trails retained at least as long as the subject records. 21 CFR 11.10(а) adds validation of systems for accuracy, ышанычлылык, эзлекле максатчан эш, һәм дөрес булмаган яки үзгәртелгән язмаларны ачыклау сәләте, шул вакытта 11.100(а) һәм 11.300(а) require each electronic signature to be unique to one individual and never reused or reassigned.
The failure mode is an electronic record that cannot show what changed, кайчан, and by whom. A revised peptide certificate of analysis without a retained prior version is exactly that gap.
|
ALCOA attribute |
Record artifact that demonstrates it |
|---|---|
|
Сыйфатлы |
Named analyst and unique credential on each entry |
|
Әр сүзнең |
Time-stamped audit trail readable across revisions |
|
Заманча |
Entry timestamp matching the action |
|
Оригиналь |
Хезмәтләр Retained first capture, not a later transcription |
|
Төгәл |
Validated system able to flag altered records |
Two limits matter. FDA’s stated scope and application of Part 11 says the agency will narrowly interpret the rule, and where computers merely generate paper printouts meeting all record requirements, the paper record is the regulated record. Partлеш 11 also does not reach every electronic file a CRO holds.
The cross-reference walk-through: one lot, five hops
A minimum connected data package for a peptide project is only as strong as the links between its records, so the practical test is whether one lot number can be followed through five hops without a dead end.
Take a lot released by MOL Changes. The lot number on the certificate of analysis points to the RP-HPLC chromatogram, which names the method ID it was run under. That method ID points to the stability protocol the same lot entered, and the protocol points to the assay record that used it. Each hop is checkable. None of them, үзеннән-үзе, tells you the material is fit for your experiment.
The solubility record shows where the chain stops being about release. It is run to find optimal solvent conditions, can be customized to the solvents your experiment uses, with water as the first choice, and it answers a formulation question rather than a release question. A reviewer can conclude the lot dissolves under the tested conditions. They cannot conclude it will dissolve in a different buffer, at a different concentration, or after freeze-thaw.
Reviewer checklist and next steps
Use this as a reusable filter for any supplier, not just the one you are evaluating now. It condenses the peptide CRO documentation requirements into the questions a reviewer actually asks before accepting a package.
-
Is the certificate of analysis batch-specific, or a generic template?
-
Does the supplier state its chromatography data system’s audit-trail policy?
-
Will they release a representative batch record for a comparable sequence length and hydrophobicity, including the impurity profile?
-
Is there a scale-up or Кибет чагыштыру отчеты?
-
Is the method-transfer record complete?
-
Do you receive the actual chromatogram and mass spectrum, or only a summary table?
-
For a modified sequence, is there a prior batch record carrying the same modification? Турында
-
Do the records follow MHRA ALCOA principles, and does the supplier hold an ISO 13485 яки ISO 9001 сертификат?
A due-diligence playbook for peptide CRO selection turns on one decision rule: if any of these seven records is missing and the supplier will not commit to a date for it, treat the package as unverified and price the risk into your timeline. The checklist is reusable across suppliers, which is what makes switching less costly than it looks.
MOL Changes ships a full certificate of analysis with chromatograms, downloadable from each product page and retrievable via a peptide-specific QR code, and it has a commercial interest in peptide quality standards. If you want to test the checklist against a live package, request a sample data package or talk to an analytical expert.
[CTA: Request a sample data package]
Еш бирелә торган сораулар
What must a peptide certificate of analysis contain?
A usable peptide certificate of analysis identifies the lot, names the method behind each result, and reports purity, үзенчәлек, and content as separate values rather than one headline figure. A reviewer needs the lot number, method identifiers, the chromatogram and mass spectrum, һәм спецификация һәр бәягә каршы хөкем ителә. A package reduced to one purity percentage leaves the receiving lab unable to confirm what was measured.
Can purity percentages be compared across vendors?
Only when the chromatographic method is comparable, because a purity figure is a property of the method as much as of the peptide. Two vendors reporting 98% may be running different columns, gradients, һәм дулкын озынлыкларын ачыклау, and a method that resolves fewer impurities reports a higher number for the same material. Treat a percentage without its method parameters as unverifiable rather than comparable.
How much stability data does a discovery-stage peptide program need?
Enough to cover the interval and conditions the material will actually meet, and no more. Early discovery work rarely justifies a full ICH Q1A(R2) bracketed study, but it does justify knowing whether the peptide survives the buffer, температура, and handling it meets during the assay. Where sterility is part of the specification, the USP <71> sterility test sets a concrete floor: media are incubated for not less than 14 көннәр, with fluid thioglycollate medium at 32.5 ± 2.5 °C and soybean-casein digest medium at 22.5 ± 2.5 ° C., and no evidence of microbial growth means the product complies.
Can you request raw chromatograms and mass spectra?
Әйе, and for any lot you intend to rely on, you should. A summary purity figure is a conclusion; the chromatogram shows peak shape, baseline, and whether a leading shoulder was integrated into the main peak or excluded from it. Some vendors publish full chromatograms as standard practice, others provide them on request. Ничек кенә булмасын, the request belongs in the documentation requirements you set before the first order, not after a batch fails.
How long should a peptide data package be retained?
Retain it for as long as any result derived from that lot may need defending, which in practice means the life of the program plus the retention period your quality system requires. Stability conclusions, assay qualifications, and any submission citing the material all trace back to the original records, so a package that cannot be retrieved later is effectively a package that never existed. Confirm the retention commitment as part of your peptide CRO documentation requirements rather than assuming it.
Йомгаклау
The minimum connected data package for a peptide project is defined by re-runnability, not by volume: if a receiving lab cannot repeat your result from the records alone, the package is incomplete no matter how many pages it runs to. That test sorts the five claims cleanly. Чисталык, үзенчәлек, эчтәлеге, stability and assay suitability each need their own record, and each record carries a stated limit on what it does not prove. The cross-reference chain holds them together, from lot number through chromatogram and method ID to the stability protocol, so a reviewer can follow one hop at a time instead of reconstructing your project from memory.
The standards layer underneath is stable. I Q2(R2) governs how the analytical procedures are validated, I Q1A(R2) and Q5C set the stability expectations, USP monographs fix the compendial tests, һәм 21 CFR өлеше 11 covers the audit trail. Vendor practice is not stable, and that gap is where packages fail review.
So the next step is concrete. Take the last certificate of analysis your current supplier sent you, pick one lot, and try to walk the five hops without emailing anyone. Where the chain breaks, that is the question to put to your supplier in writing.