"펩타이드 프로젝트를 위한 최소 연결 데이터 패키지"가 실제로 의미하는 것
펩타이드 합성 정의: 펩타이드 프로젝트에 대한 최소 연결 데이터 패키지는 두 번째 실험실에서 결과를 재현하고 검토자가 결정을 방어할 수 있는 가장 작은 상호 참조 기록 세트입니다..
연결된 것은 완전한 것과 같지 않다. 완전한 패키지에 모든 문서가 들어있습니다.; 연결된 문서는 각 문서가 다른 문서를 가리키도록 만듭니다., 그래서 많은 레코드에서는 크로마토그램에 이름을 붙입니다., 크로마토그램은 해당 방법의 이름을 지정합니다., 메소드는 실행된 표준의 이름을 지정합니다.. MOL 변경 사항은 재실행 시 재현성 테스트를 작성합니다.: 패키지가 질문에 대한 답변을 제공하지만 서면 기록만으로는 두 번째 랩에서 다시 실행할 수 없는 경우, 과정보다는 결과를 기록한다. 이것이 바로 이 가이드가 오류를 방지하기 위해 만들어진 오류 모드입니다..
실제로 연결된 기록이 어떤 모습인지는 완전한 증거 패키지의 실제 사례에서 설명됩니다.: 불순물 이론적 근거가 있는 합성 경로, 동일한 로트의 출시 및 안정성 데이터, 실제 결과와 함께 허용 기준이 포함된 분석법 이전 보고서, 원시 데이터를 검색 가능한 상태로 유지하는 인증서.
5개의 주장이 그 구조 안에 있으며 결코 서로를 대체해서는 안 됩니다.: 청정, 신원, 콘텐츠, 안정, 및 분석 적합성. 이 가이드의 나머지 부분에서는 한 번에 하나씩 설명합니다..
패키지가 별도로 유지해야 하는 5가지 주장
청정, 신원, 콘텐츠, 안정성과 분석 적합성은 다섯 가지 다른 주장입니다., 그리고 각각은 자신의 증거가 필요합니다. 이를 혼동하는 것이 가장 일반적인 검토 실패입니다., 일반적으로 하나의 순도 수치를 전체 특성화로 읽는 것으로 시작됩니다..
펩타이드 취급에 대해 발표된 방법론적 권장 사항이 관례를 설정합니다.: 정제된 연구 펩타이드는 일반적으로 다음보다 큰 값으로 지정됩니다. 95% 화학적 순도, 분취용 RP-HPLC 정제 및 세 가지 QC 단계로 지원됩니다., 순도 백분율 분석용 RP-HPLC, 질량 및 서열 확인을 위한 MALDI/ESI-MS 또는 MS/MS, 농도 및 조성 정확도를 위한 아미노산 분석 (세대를 위한 권고사항, 부량, 펩타이드의 보관 및 취급). 별거는 집 취향이 아니다. 나는 Q2(R2), 6월부터 2024, 이름 분석 / 효능, 청정, 별도의 분석 목적으로 불순물과 정체 확인, 각각은 고유한 검증 기대치를 가지고 있습니다., 그리고 USP 펩타이드 참조 표준 및 불순물 표준 공정 관련 불순물과 분해 불순물을 하나의 순도 수치가 아닌 별도의 범주로 처리.
펩타이드 순도와 동일성의 차이는 검토자가 먼저 테스트하는 것입니다., 왜냐하면 두 가지가 서로 다른 도구로 측정되고 서로 다른 질문에 대답하기 때문입니다..
|
Cl 합성 펩티드 목표 |
증거요건 |
증명하지 못하는 것 |
|---|---|---|
|
청정 |
분석적 RP-HPLC 면적 백분율 |
직교 증거 없이 동시 용출 잘림과 산화를 구별할 수 없음 |
|
신원 |
이론질량 대비 질량 확인 |
순도나 내용을 확립하지 않습니다. |
|
콘텐츠 / 순 펩티드 |
아미노산 분석, 반대이온과 물이 포함되어 있다 |
면적 비율은 콘텐츠 값이 아닙니다. |
|
안정 |
명시된 조건에서의 시점 데이터 |
출시 당시의 신원이나 콘텐츠에 대해서는 아무 말도 하지 않습니다. |
|
분석 적합성 |
방법별 적격성 평가 |
다른 분석 형식으로 전환되지 않습니다. |
제한은 테이블보다 더 중요합니다.. 순도 면적 백분율은 함량 또는 순 펩타이드 값이 아닙니다., 왜냐하면 반대이온, 크로마토그램에서 물과 잔류 용매 질량은 고려되지 않습니다.. 많이 읽을 수 있다 97% 순수하고 라벨에 표시된 것보다 훨씬 적은 양의 펩타이드를 전달합니다..
시퀀스 기록 및 수정 내역
시퀀스 레코드는 패키지의 기본 키입니다.. 다른 모든 기록 (크로마토그램, 질량 스펙트럼, 안정성 프로토콜, 분석 결과) 특정 버전에 대해서만 해석 가능, 따라서 해당 레코드가 의미를 갖기 전에 버전 관리를 거쳐야 합니다..
리뷰어가 이를 사용하려면 6개의 필드가 필요합니다.: 시퀀스 버전, N- 및 C 말단 주석, 변형 부위 및 화학, 반대이온 형태, 및 로트 ID. 수정 필드에 가장 많은 가중치가 적용됩니다., 변형된 잔여물은 패키지의 추적성이 가장 자주 상실되는 부분이기 때문입니다.. 펩타이드 CRO 선택을 위한 실사 플레이북은 이전 배치 기록 또는 동일한 변형이 포함된 시퀀스에 대한 CoA를 처리합니다., 수정 위치를 확인하는 데 사용되는 분석 방법, 문서 요구 목록의 일부로, 선택적인 추가사항이 아닌.
수정에 미리 만들어진 분석 경로가 없는 경우, 참조 표준이 작업을 수행합니다.. USP는 펩타이드 API 참조 표준을 공급합니다., 관련 불순물 참조 표준품 및 분석 참조 자료, 또한 Oxytocin 식별 참조 표준 및 Goserelin 시스템 적합성 혼합물과 같은 시스템 적합성 표준, 방법을 개발 및 검증하고 원료의약품 및 제품 배치를 관리하는 데 사용할 수 있습니다. (USP 생물학제, 펩타이드 표준). 펩타이드 생산
실패 모드는 변경할 수 없는 버전 추적이 없는 시퀀스 편집입니다.. 재현 불가능한 실험의 가장 흔한 원인입니다.: 많은 것이 진짜다, 데이터는 진짜야, 어떤 분자가 만들어졌는지 아무도 증명할 수 없습니다.
순도 및 신원 기록: 크로마토그램과 질량 스펙트럼이 실제로 증명하는 것
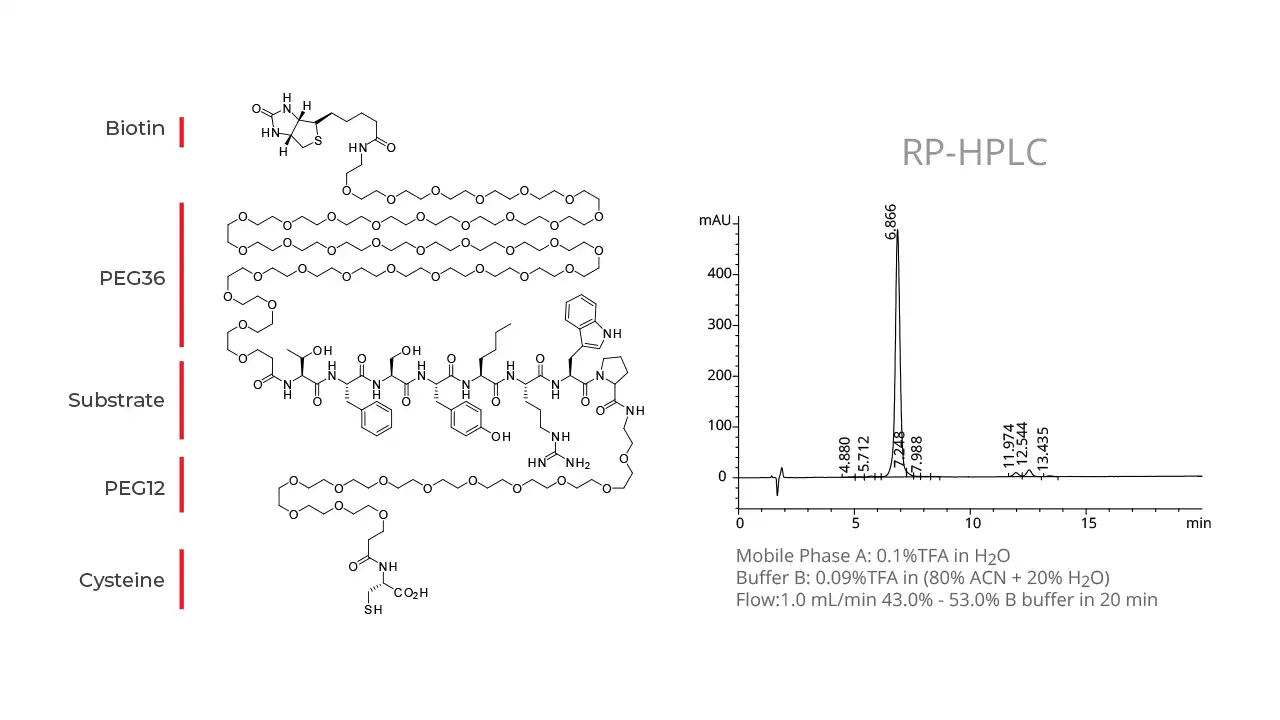
RP-HPLC는 상대 순도를 확립합니다.; ESI-MS는 신원을 확립합니다. 어느 쪽도 다른 쪽을 확립하지 않습니다., 그리고 그것들을 상호 교환 가능한 것으로 취급하는 패키지는 리뷰어가 두 질문 중 하나에도 대답할 수 없게 만듭니다..
구별은 사설이 아닙니다.. 나는 Q2(R2), 6월부터 2024, 2분기 교체됨(R1) 분석/역가를 치료합니다., 청정, 별도의 분석 목적으로 불순물과 정체 확인, 각각은 고유한 검증 기대치를 가지고 있습니다.. 무형유산 Q2(R2) FDA가 발행한 검증 지침 3월 2024 동일한 구조를 가지고 있습니다. 따라서 리뷰어의 첫 번째 질문은 "HPLC 추적이 있습니까?"가 아니라 "이 방법은 어떤 목적에 도움이 됩니까?"입니다., 그리고 그것이 그 목적으로 검증되었는지.”
크로마토그램의 경우, 중요한 필드는 열과 차원입니다., 구배, 검출 파장, 참조 표준, 및 메소드 ID. 선두적인 어깨 0.6 RRT는 해당 매개변수에 대해서만 해석 가능합니다.: 그라데이션 없이, 유지 시간은 아무 의미가 없습니다.
펩타이드 CRO 소싱 실무 분석 구매자는 HPLC 및 대량 사양 데이터를 추가 비용이 아닌 표준으로 기대해야 한다고 명시, 요약이 아닌 원시 QC 데이터를 요청할 수 있어야 합니다..
여기에는 두 가지 한계가 속합니다.. 요약표는 크로마토그램이 아닙니다.. 그리고 질량 스펙트럼은 질량을 확인합니다, 시퀀스 위치가 아님: 삭제 및 대체는 동일한 명목 질량을 반환할 수 있습니다., 따라서 MS만으로는 시퀀스가 다른 부분을 알 수 없습니다..
콘텐츠, 반대이온과 물질수지
콘텐츠, 또는 순 펩티드, 물질수지 값이다: 무게를 잰 분말 중 실제로 얼마나 많은 펩타이드가 들어 있는지 명시되어 있습니다., 반대이온 후, 물과 잔류 용매를 뺍니다.. 완전해 보이는 패키지에서 가장 자주 누락되는 주장입니다., HPLC 면적 비율은 이를 대신할 수 없습니다.. 면적 백분율은 한 파장에서 UV 흡수 피크 면적의 비율을 측정합니다.; 피크가 나타내는 물질의 질량에 대해서는 아무 말도하지 않습니다..
순수성 전용 인증서를 읽는 검토자는 어쨌든 산술을 수행합니다.. 많이 신고된 경우 95% 면적별 순도 및 순 펩타이드 함량은 70%, 그럼 10 mg 무게 측정은 대략적으로 전달됩니다. 7 mg의 펩티드, 모든 다운스트림 몰 계산은 그 차이를 상속받습니다.. 반대이온은 일반적으로 가장 큰 단일 용어입니다.: 아세테이트와 트리플루오로아세테이트는 모두 펩타이드가 아닌 질량을 추가합니다..
아미노산 분석은 농도 및 구성 정확도에 대한 참조 접근 방식입니다., 크로마토그래피 반응에서 추론하는 대신 직접적으로 펩타이드 질량을 정량화하기 때문입니다. (제약 분석 저널, 2016).
핵심 내용: 면적 백분율에 따른 순도와 질량 균형에 따른 함량이 다양한 질문에 답합니다.. 첫 번째 항목만 보고하는 패키지에서는 검토자가 두 번째 항목을 추측하게 됩니다..
패키지가 콘텐츠를 보고하지만 그 뒤에 있는 방법은 보고하지 않는 경우, 번호는 확인불가, 이를 전달해야 하는 인증서 섹션은 비어 있습니다..
안정성 기록: 설계, 조건 및 시점
안정성 결과 테이블은 이를 생성한 프로토콜이 동일한 패키지로 이동하는 경우에만 해석 가능합니다.. 프로토콜 없이, a receiving lab cannot tell whether the study applies to its own storage and handling conditions.
The required contents of a stability protocol are set out in Health Canada’s guidance on stability, 섹션 52 에게 62 (2025-09-04). A peptide stability data package should carry, 해당하는 경우: product identification and master-document references; container and closure system type and size, plus total container count; lot or batch code with manufacture date and proposed expiry; sample size and test intervals per parameter; storage conditions with tolerances and how those tolerances are maintained; sample orientation reflecting worst case; specifications with acceptance criteria per parameter; the method of data evaluation, including statistical analysis or trending; and the report format.
These fields sit inside a framework rather than replacing it. 그만큼 current Q1A(R2) stability framework dates to November 2003, 그리고 the Q5C expectations for peptide-like products (칠월 1996) state that products whose active components are typically proteins or polypeptides are particularly sensitive to temperature, 산화, 빛, ionic content and shear, so stringent storage conditions are usually necessary. Q5C does not supersede Q1A(R2); the two apply together. The field list above is a general stability-protocol requirement, not a peptide-specific or FDA-specific one.
분석 적합성 및 결과를 전송할 수 있는 기록
Assay suitability belongs to the method in the matrix, not to the peptide. A lot that reads clean by RP-HPLC can still fail a cell-based potency assay, because the assay measures what the peptide does in a buffer, a cell line and a plate, and those conditions are part of the result. The records that make a potency number transferable are therefore the format, the matrix, the controls and the acceptance criteria, not just the value.
For conjugate-type programs the bar is higher. ChemExpress sets out the analytical demands for peptide drug conjugate programs as LC-MS/MS for structural confirmation, SEC-MALS for aggregation, HPLC-UV/MS for drug-to-peptide ratio and free payload, peptide mapping for modification sites, and bioassays for potency, with methods validated per ICH guidelines. The same source states that without conjugation-site, drug-to-peptide ratio and aggregation characterization, “you cannot file an IND.”
Where a discovery-stage package may stop short: if the program is not yet filing, a documented method with matrix and controls can be enough, provided the record says which of those characterizations are still open.
기록, 감사 추적 및 패키지를 방어 가능하게 만드는 요소
A package is defensible when its electronic records meet the same integrity expectations as paper ones, and Part 11 states those expectations directly. 21 CFR 11.10(이자형) and the audit-trail requirement calls for secure, 컴퓨터 생성, time-stamped audit trails that independently record the date and time of operator entries and actions creating, 수정, or deleting electronic records, with changes never obscuring previously recorded information and trails retained at least as long as the subject records. 21 CFR 11.10(에이) adds validation of systems for accuracy, 신뢰할 수 있음, consistent intended performance, and the ability to discern invalid or altered records, ~하는 동안 11.100(에이) 그리고 11.300(에이) require each electronic signature to be unique to one individual and never reused or reassigned.
The failure mode is an electronic record that cannot show what changed, 언제, and by whom. A revised peptide certificate of analysis without a retained prior version is exactly that gap.
|
ALCOA attribute |
Record artifact that demonstrates it |
|---|---|
|
귀속 |
Named analyst and unique credential on each entry |
|
읽기 쉬운 |
Time-stamped audit trail readable across revisions |
|
동시대의 |
Entry timestamp matching the action |
|
원래의 |
서비스 Retained first capture, not a later transcription |
|
정확한 |
Validated system able to flag altered records |
Two limits matter. FDA’s stated scope and application of Part 11 says the agency will narrowly interpret the rule, and where computers merely generate paper printouts meeting all record requirements, the paper record is the regulated record. 부분 11 also does not reach every electronic file a CRO holds.
상호 참조 연습: 한 몫, 다섯 개의 홉
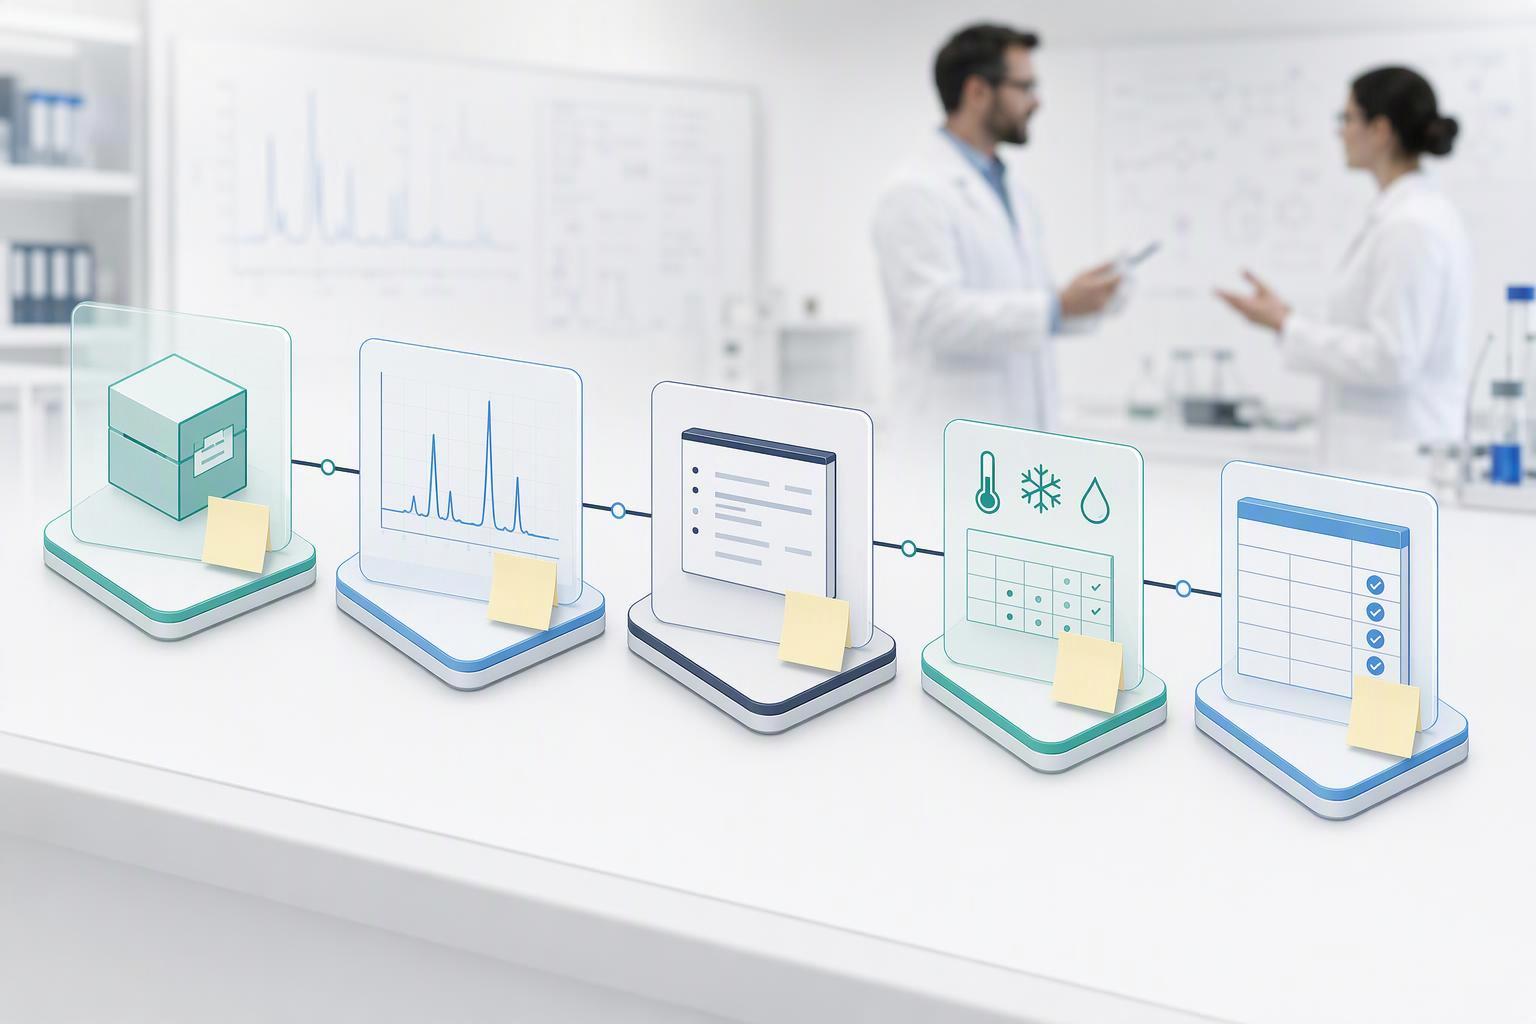
A minimum connected data package for a peptide project is only as strong as the links between its records, so the practical test is whether one lot number can be followed through five hops without a dead end.
Take a lot released by MOL Changes. The lot number on the certificate of analysis points to the RP-HPLC chromatogram, which names the method ID it was run under. That method ID points to the stability protocol the same lot entered, and the protocol points to the assay record that used it. Each hop is checkable. None of them, 그 자체로, tells you the material is fit for your experiment.
The solubility record shows where the chain stops being about release. It is run to find optimal solvent conditions, can be customized to the solvents your experiment uses, with water as the first choice, and it answers a formulation question rather than a release question. A reviewer can conclude the lot dissolves under the tested conditions. They cannot conclude it will dissolve in a different buffer, at a different concentration, or after freeze-thaw.
리뷰어 체크리스트 및 다음 단계
Use this as a reusable filter for any supplier, not just the one you are evaluating now. It condenses the peptide CRO documentation requirements into the questions a reviewer actually asks before accepting a package.
-
Is the certificate of analysis batch-specific, or a generic template?
-
Does the supplier state its chromatography data system’s audit-trail policy?
-
Will they release a representative batch record for a comparable sequence length and hydrophobicity, including the impurity profile?
-
Is there a scale-up or 가게 comparability report?
-
Is the method-transfer record complete?
-
Do you receive the actual chromatogram and mass spectrum, or only a summary table?
-
For a modified sequence, is there a prior batch record carrying the same modification? 에 대한
-
Do the records follow MHRA ALCOA principles, and does the supplier hold an ISO 13485 또는 ISO 9001 certificate?
A due-diligence playbook for peptide CRO selection turns on one decision rule: if any of these seven records is missing and the supplier will not commit to a date for it, treat the package as unverified and price the risk into your timeline. The checklist is reusable across suppliers, which is what makes switching less costly than it looks.
MOL Changes ships a full certificate of analysis with chromatograms, downloadable from each product page and retrievable via a peptide-specific QR code, and it has a commercial interest in peptide quality standards. If you want to test the checklist against a live package, request a sample data package or talk to an analytical expert.
[CTA: Request a sample data package]
자주 묻는 질문
펩타이드 분석 증명서에는 무엇이 포함되어야 합니까??
A usable peptide certificate of analysis identifies the lot, names the method behind each result, and reports purity, 신원, and content as separate values rather than one headline figure. A reviewer needs the lot number, method identifiers, the chromatogram and mass spectrum, and the specification each value is judged against. A package reduced to one purity percentage leaves the receiving lab unable to confirm what was measured.
공급업체 간에 순도 비율을 비교할 수 있습니까??
Only when the chromatographic method is comparable, because a purity figure is a property of the method as much as of the peptide. Two vendors reporting 98% may be running different columns, gradients, 및 검출 파장, and a method that resolves fewer impurities reports a higher number for the same material. Treat a percentage without its method parameters as unverifiable rather than comparable.
발견 단계의 펩타이드 프로그램에는 얼마나 많은 안정성 데이터가 필요합니까??
Enough to cover the interval and conditions the material will actually meet, and no more. Early discovery work rarely justifies a full ICH Q1A(R2) bracketed study, but it does justify knowing whether the peptide survives the buffer, 온도, and handling it meets during the assay. Where sterility is part of the specification, the USP <71> sterility test sets a concrete floor: media are incubated for not less than 14 날, with fluid thioglycollate medium at 32.5 ± 2.5 °C and soybean-casein digest medium at 22.5 ± 2.5 ℃, and no evidence of microbial growth means the product complies.
원시 크로마토그램과 질량 스펙트럼을 요청할 수 있나요??
예, and for any lot you intend to rely on, you should. A summary purity figure is a conclusion; the chromatogram shows peak shape, baseline, and whether a leading shoulder was integrated into the main peak or excluded from it. Some vendors publish full chromatograms as standard practice, others provide them on request. 어느 쪽이든, the request belongs in the documentation requirements you set before the first order, 배치 실패 이후가 아님.
펩타이드 데이터 패키지를 얼마나 오래 보관해야 합니까??
Retain it for as long as any result derived from that lot may need defending, which in practice means the life of the program plus the retention period your quality system requires. Stability conclusions, assay qualifications, and any submission citing the material all trace back to the original records, so a package that cannot be retrieved later is effectively a package that never existed. Confirm the retention commitment as part of your peptide CRO documentation requirements rather than assuming it.
결론
The minimum connected data package for a peptide project is defined by re-runnability, not by volume: if a receiving lab cannot repeat your result from the records alone, the package is incomplete no matter how many pages it runs to. That test sorts the five claims cleanly. 청정, 신원, 콘텐츠, stability and assay suitability each need their own record, and each record carries a stated limit on what it does not prove. The cross-reference chain holds them together, from lot number through chromatogram and method ID to the stability protocol, so a reviewer can follow one hop at a time instead of reconstructing your project from memory.
The standards layer underneath is stable. 나는 Q2(R2) governs how the analytical procedures are validated, 나는 Q1A(R2) and Q5C set the stability expectations, USP monographs fix the compendial tests, 그리고 21 CFR 부분 11 covers the audit trail. Vendor practice is not stable, and that gap is where packages fail review.
So the next step is concrete. Take the last certificate of analysis your current supplier sent you, pick one lot, and try to walk the five hops without emailing anyone. Where the chain breaks, that is the question to put to your supplier in writing.