What the FDA NAM Rule Actually Changes for a Peptide CMC Package
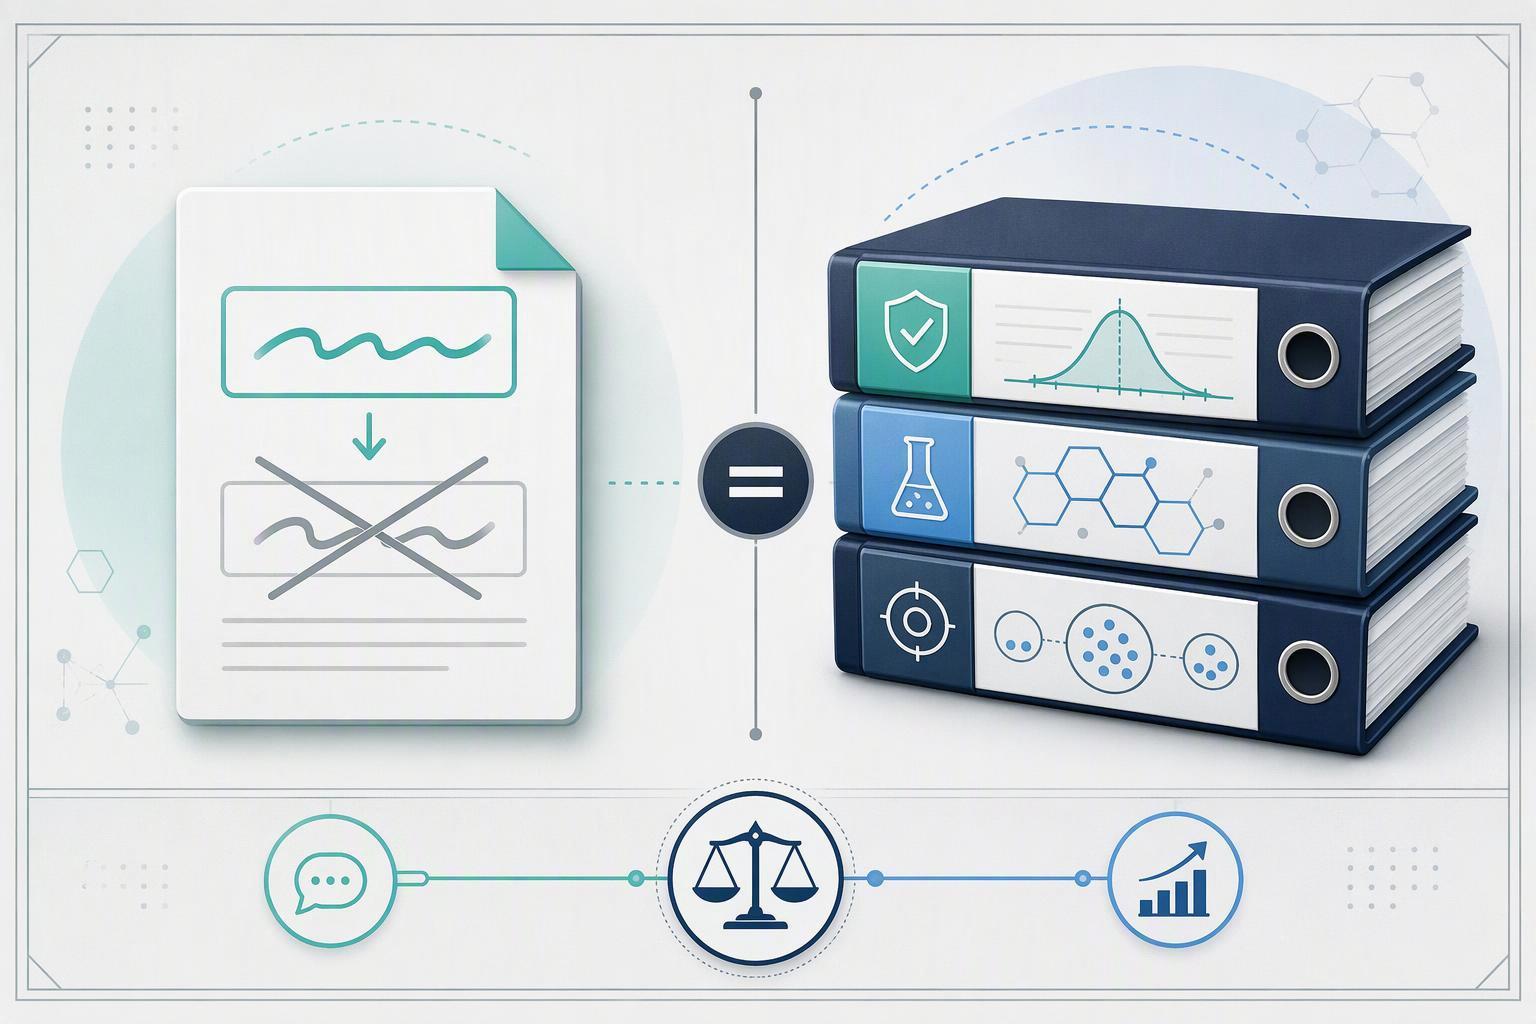
A peptide CMC package is the analytical and manufacturing evidence that defines what you dosed, how pure it was, and how it behaved over time. The FDA NAM rule is the terminology change that moved the agency’s nonclinical vocabulary from “preclinical” to “nonclinical,” and the two are now discussed together because a nonclinical study that no longer rests on a species-based model leans harder on that analytical record.
The NAM rule itself is narrower than most summaries suggest. The FDA Modernization Act 2.0 replaced “preclinical” with “nonclinical” in the statute, Thiab FDA 2026 nonclinical-testing terminology rule is regulatory clean-up that aligns the agency’s language with that shift. It does not eliminate or prohibit animal studies, change evidentiary standards, or impose new costs or requirements. If you were waiting to learn whether your filing obligations changed, they did not.
⚠️ Warning: Do not read the rule as a ban. The language changed; the evidence bar did not. What changed is the vocabulary FDA uses and the wording that implied animal testing was the only acceptable route. What stayed untouched is everything that determines whether your submission survives review: evidentiary standards, cost and requirement burden, and the identity, purity, and potency evidence a test article must carry.
Why the CMC Package Became the Load-Bearing Part of the Preclinical Story
This is the shift you are probably feeling. When a nonclinical study no longer rests on a species-based model, the analytical record of what was actually dosed carries the interpretive weight that the animal model used to share.
FDA has described this transition as strategic and stepwise, backed by validation frameworks and success metrics, and its 2026 draft NAM guidance supplies a validation framework and regulatory expectations for sponsors making the move. Lub koom haum tsis thov kom koj tso tseg pov thawj. Nws thov kom koj qhia, kev tshuaj xyuas, tias cov ntawv kuaj hauv koj txoj kev tshawb fawb yog qhov koj hais tias nws yog.
ISTAND hloov mus rau ib qho kev kawm ntawv pov thawj tas mus li qhia tau tias qhov ntawd tau mus deb npaum li cas: yim qhov kev xa ntawv tau txais, faib ua peb AI-based, ob qho kev nyab xeeb ua ntej tsis yog tsiaj, ob qho raws li cov ntaub so ntswg, thiab ib txoj kev txheeb xyuas lej.
Peptide Characterization: Proving the Test Article Is What You Say It Is
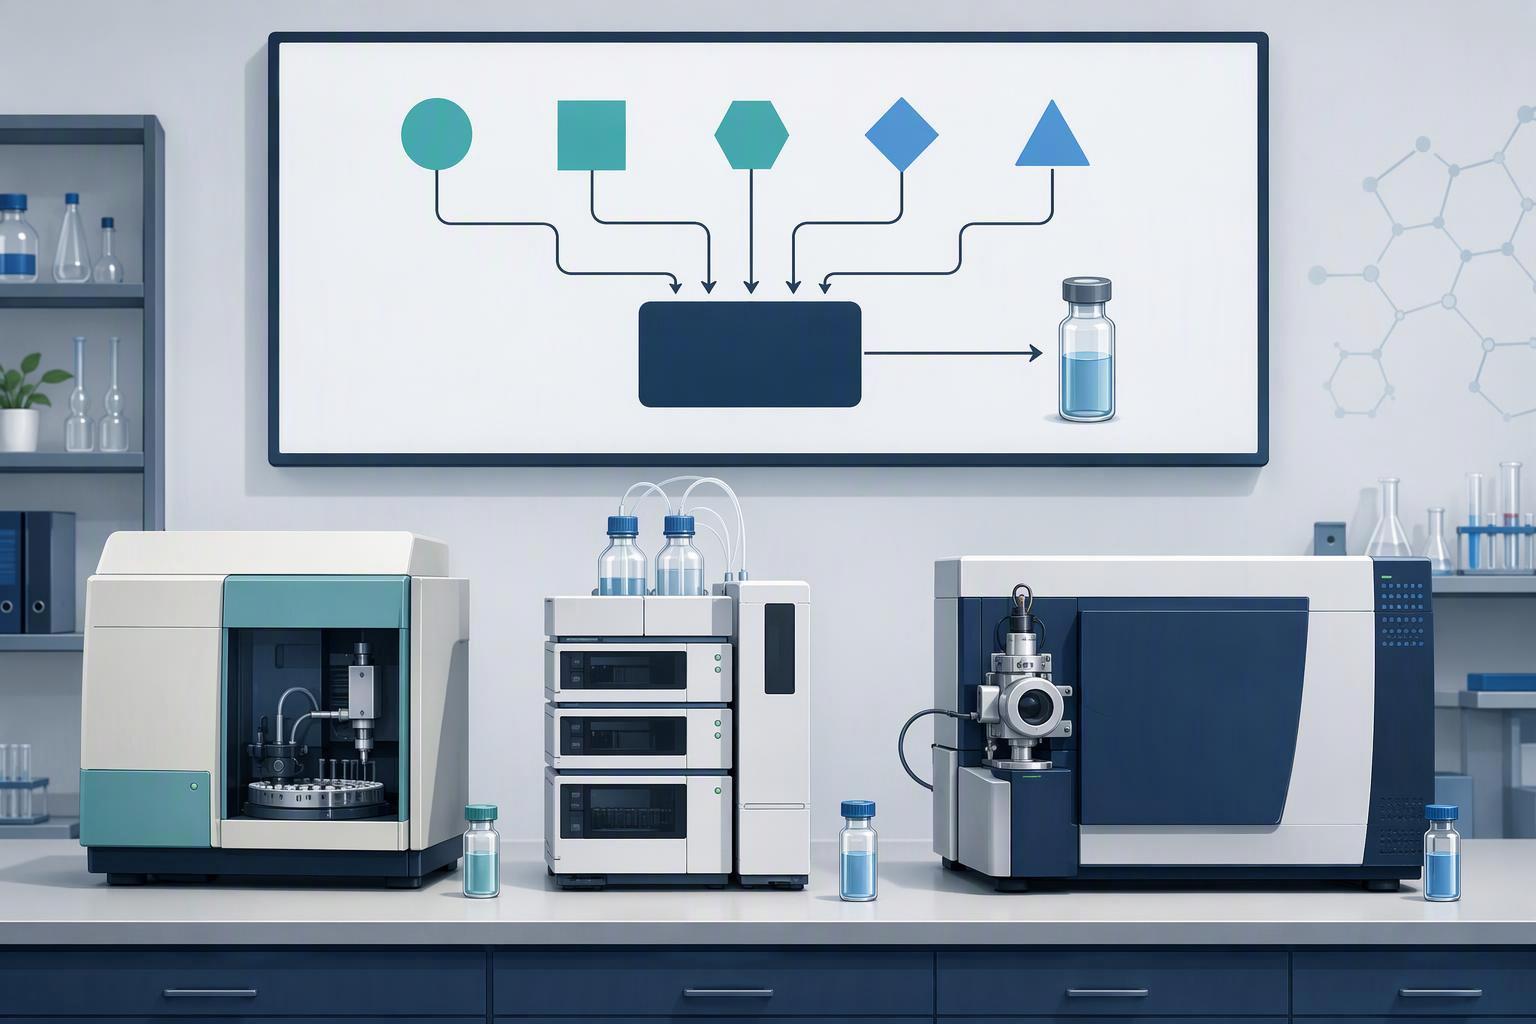
Kev piav qhia yog thawj vim txhua yam tom qab tau txais nws qhov tsis paub tseeb. Yog koj tsis tuaj yeem qhia tau tias cov khoom siv hauv kev tshawb fawb NAM yog tib molecule li koj tus neeg tsim kho, cov impurity, kev ruaj ntseg, thiab cov ntaub ntawv assay uas tsim saum nws piav qhia ib qho tshuaj uas koj tsis tau piav qhia tiag tiag.
Identity for a peptide rests on orthogonal methods rather than a single technique. Mass spectrometry and LC-MS/MS establish molecular mass, amino acid analysis including chiral amino acid analysis confirms composition and stereochemistry, peptide mapping and NMR interrogate sequence and structure, and counterion identity is typically resolved by ion chromatography or reversed-phase HPLC. These are the method families named in FDA’s stated CMC expectations for therapeutic peptides, and the pairing matters: two methods that share the same failure mode confirm nothing.
For peptide characterization for CMC, the practical minimum is two orthogonal methods for identity and for each critical quality attribute, not one method run twice. The “so what” is representativeness, and it has to survive change. A post-manufacturing change, whether a new supplier, scale, or purification step, obliges you to re-run the same method set on the changed batch and show the identity conclusion still holds. A characterization package that only describes the original batch does not cover the material a later study will use.
Peptide Impurity Profiling: Why There Is No Universal Threshold
There is no single impurity limit a peptide program has to hit. Limits are set case by case from batch history, stability data and toxicology findings, and the familiar 0.1% figure is a reporting convention rather than a safety threshold. The FDA’s 0.10% to 0.5% impurity window for new peptide-related impurities exists because immunogenicity risk rises with impurities in that range, thiab qhov 0.5% upper bound is justified on the grounds that it is consistent with the small amount of unspecified peptide-related impurities observed in finished peptide products.
The mechanism behind that risk-based approach is a coverage gap. Peptide-related impurities sit outside full ICH Q3A coverage, so a program cannot simply point to a default qualification threshold and stop there. Justification has to be built from the program’s own data.
Process-related impurities and degradation products also need separate arguments. Process-related impurities come from synthesis, purification and handling, so the control case rests on the manufacturing process and its clearance steps. Degradation products form in the drug substance or product over time, so the case rests on stability data and the analytical methods that detect them. One control strategy rarely answers both.
Stability Testing: What Makes a Method Stability-Indicating
A stability-indicating assay for peptides is an analytical method that can measure the active peptide in the presence of its degradation products, excipients, and process impurities, and it earns that description only when forced degradation has demonstrated the separation. The distinction matters because a method that resolves the main peak cleanly tells you nothing about whether a degradant is hiding underneath it.
Forced degradation is how you settle that question before a shelf-life claim is made. Expose the peptide to acid, base, peroxide, heat, light, and humidity stress, then confirm that degradation peaks are resolved from the main peak and that peak purity is maintained across the stressed samples. If a degradant co-elutes with the peptide, the assay will report a purity that the sample does not have, and any stability trend built on it is unreliable.
Validation then follows the framework of the validation characteristics in ICH Q2(R2), which sets out accuracy, precision, specificity, detection limit, quantitation limit, linearity, and range, adopted with a legal effective date of 14 June 2024. Specificity carries the most weight here, because it is the characteristic that documents the forced-degradation work.
Pro Tip: Run forced degradation before you commit to a shelf-life claim, not after. Reversing that order usually means repeating the stability study once the method changes.
Where the evidence stops: ICH Q2(R2) defines the characteristics a method must demonstrate, but it does not prescribe a single forced-degradation protocol or acceptance threshold for peptides. Those are set case by case against the route of degradation your molecule actually shows.
Analytical Assay Development: Validation Characteristics That Survive Review
“Validated” means the assay has documented evidence that it performs as intended for its specific purpose, not that it passed a generic checklist. For analytical assay development for peptide programs, that evidence is organized around a defined set of validation characteristics, and each one catches a different way a peptide result can mislead you.
|
Validation characteristic |
Peptide-specific failure it catches |
|---|---|
|
a |
A method that reads pure reference standard correctly but drifts on the real test article |
|
Precision |
Cardiovascular Solid Phase Peptide Synthesis Resin Repeatability that looks acceptable within one run but collapses across days or analysts |
|
Specificity |
Related impurities or degradation products co-eluting with the main peak |
|
Linearity Cell Penetrating and range |
Quantitation extrapolated beyond the concentrations the method actually supports |
|
Limit of detection Peptide Modification and quantitation |
Trace impurities reported at levels the method cannot reliably measure |
|
Robustness |
Small, permitted changes in column, pH, or flow that silently shift results |
The four domains are one argument, not four service lines. Characterization defines what the material is, impurity profiling defines what else is present, stability defines how both change over time, and the assay is the instrument that measures all three. When a manufacturing change occurs, comparability is shown by extensive chemical, physical, and bioactivity comparisons with side-by-side analyses of old product and qualification lots of new product, using release tests plus tests directed at the impact of the change, the approach set out in FDA’s long-standing comparability expectations for post-change material (1996). An assay that cannot support that side-by-side comparison cannot support the package.
Connecting Alternative Preclinical Models to a Credible Development Package
An alternative preclinical model only carries weight when the analytical record around it can be read on its own. The CMC package is what turns a study result into something a reviewer can interpret, and comparability is what keeps that interpretation valid as a program moves.
FDA’s own framing supports this. Where a manufacturing change does not affect safety, tus kheej, purity or potency, the agency may find the products comparable, which means the analytical data, not the narrative, decides whether a batch-to-batch or post-change comparison holds (FDA guidance on comparability, tau txais rov qab 2026-06-11).
The regulatory direction is still moving. The FDA Modernization Act 3.0, S.355, had its latest action on 2025-12-17 and remains pending, so any assumption about how quickly alternative-model data becomes routine should stay hedged. The ISTAND qualification pathway is the more established route for qualifying a new approach, and it depends on exactly the kind of characterization, impurity, stability and assay evidence this guide has walked through.
Qhov tseem ceeb: A NAM study is only as credible as the analytical package that makes its test article, its impurities and its stability defensible.
Programs that treat characterization, v, stability and assay development as one interlocking record, rather than four separate workstreams, are better positioned when a reviewer asks how a change was shown not to affect the product. MOL Changes supports that kind of integrated analytical work for peptide programs.
Common Misconceptions About the NAM Rule and Peptide CMC
Four misreadings of the NAM rule keep resurfacing in peptide CMC planning, and each one can push a filing strategy in the wrong direction. Here is what the rule does and does not do. Cell Repair And Regeneration
It did not ban animal testing. The NAM rule changes what FDA expects in a submission and opens a path for qualified alternatives. It does not remove animal studies from the toolbox, and for many endpoints a well-designed in vivo study remains the most defensible evidence available. Read it as an expansion of acceptable approaches, not a prohibition.
It did not lower the evidence bar. The rule shifts where evidence comes from, not how much scrutiny it receives. A CMC package built on an alternative model faces the same expectations for identity, purity, kev ruaj ntseg, and potency as any other program, and the characterization burden can feel heavier because the model itself is newer to reviewers. Merrifield Solid Phase Synthesis
0.1% is not a safety limit. That figure is a reporting and identification threshold, not a toxicity cutoff. Impurity limits are set case by case, based on the impurity’s structure, the dose, the route, and the duration of exposure.
A NAM-qualified tool does not substitute for CMC data. Qualification speaks to whether a tool is fit for a stated context of use. N, purity, w.
Qhov tseem ceeb: s. It does not certify an assay, t.
Tools and Resources for Building the Evidence Package
s, i. s, and all four are free to read.
h. a, i. s FDA’s NAMs hub t, and the ISTAND program page explains the qualification pathway for a novel method you intend to rely on.
Analytical standards. ICH Q2(R2) is the validation framework reviewers will measure your assay against, including how detection and quantitation limits are defined.
Peptide-specific reading. A Frontiers in Immunology analysis of the 2021 peptide ANDA guidance is useful for seeing how impurity and characterization expectations have been interpreted for peptides specifically, rather than for small molecules generally.
Read these in that order. The regulatory documents tell you what the package must demonstrate; the analytical standard tells you what evidence counts.
Getting Started: The First Three Steps
Start with an inventory, not a purchase order. Before you commission any new work, list what the current package already proves about the test article: tus kheej, purity, and the method used to establish each. Feem ntau cov pob preclinical uas tsim raws li ib qho qauv txawv twb muaj ntau cov ntaub ntawv siv tau tshaj li lawv cov kws sau ntawv paub. Cov ntawv teev qhia koj tias qhov twg ntawm plaub qhov chaw pov thawj (kev piav qhia, v, kev ruaj ntseg, kev tsim tshuaj ntsuam) uas tiag tiag tsis muaj thiab qhov twg tsuas xav tau kev sau ntawv zoo dua.
Tom qab ntawd muab txhua qhov chaw rau ib qho chaw tshwj xeeb. Ib qho yuam kev synthesis tshwm sim ua qhov chaw piav qhia. Ib qho khoom degradation uas koj tsis tuaj yeem txheeb xyuas yog qhov chaw impurity-profiling. Ib txoj kev uas tsis tuaj yeem cais peptide tag nrho ntawm nws cov khoom tawg yog qhov chaw ruaj khov. Kev hu domain ua ntej npe kev sim ua rau txoj haujlwm tsom mus rau thiab tiv thaiv kev rov ua dua txhua yam vim qhov txiaj ntsig thawj zaug tsis meej.
Kev ua haujlwm degradation ua ntej kev cog lus txog lub neej khaws cia. Development teams that begin characterization early tend to surface synthesis errors, degradation risks, and impurity patterns before scale-up, which reduces rework later. Forced degradation is cheap at small scale and expensive to retrofit once a stability protocol is running.
If you want a second read on where your package stands, our analytical team can walk through the characterization and impurity data with you. MOL Changes provides peptide analytical services, and this article is technical information rather than regulatory or legal advice.
Cov Lus Nug Nquag Nug
Does the FDA NAM rule change what a peptide CMC package has to contain?
PM. The rule changes how much weight an alternative model can carry in the preclinical story, not the chemistry, manufacturing, and controls evidence that sits underneath it. Characterization, v, kev ruaj ntseg, and assay data remain the record that shows the test article was what the study says it was.
Yog 0.1% an impurity limit for peptides?
PM. It is a reporting and identification threshold, not a pass or fail line. Reporting thresholds sit in the pharmacopeial general chapters, while the actual limit for a given impurity is set case by case from toxicology, route, dose, and duration. A peptide can fail review well below 0.1% if the impurity is genotoxic or immunogenic.
What makes a method stability-indicating for peptides?
A stability-indicating method separates the parent peptide from its degradation products, so a change in the peak reflects real loss rather than assay drift. Stress the material first (heat, light, pH, oxidation) and confirm the method resolves what that stress produces. If forced-degradation peaks co-elute with the parent, the method cannot support a shelf-life claim.
How many orthogonal methods does a peptide program need?
Enough that no single method carries the whole identity claim. Mass confirmation plus amino acid analysis plus sequencing covers most synthetic peptides; add chromatographic purity and, where relevant, chiral or counterion methods. The number follows the molecule’s risk profile, not a fixed count.
Can a NAM-qualified tool replace CMC data?
PM. A qualified alternative model replaces an animal study, not the characterization work that defines the test article. Regulators still expect the CMC package to establish identity, purity, strength, and stability for the material dosed in that model.
What changes in the CMC package after a manufacturing change?
Everything tied to the changed step gets re-examined, and comparability is the question. Run the same characterization, impurity, and stability methods against pre- and post-change material and show the profiles match within defined limits. A process change that shifts impurity profile or polymorph form can invalidate earlier toxicology assumptions.
Xaus lus
The FDA NAM rule changed the vocabulary of your preclinical filing, not the evidence bar, so the peptide CMC package under the FDA NAM rule is still where a NAM-based story is won or lost. Characterization proves the test article is what you say it is. Impurity profiling shows you control what else is in it, with limits set case by case rather than by one universal threshold. Stability testing only counts when the method is stability-indicating, meaning it separates the degradants it is meant to detect. Assay development holds the package together when its validation characteristics survive review.
The standards track keeps moving underneath all four. ICH M15 reached its final version on 2026-06-03, which means the framework you build against today may need revisiting sooner than your filing timeline assumes.
One action follows from that. Pull your current CMC package and check each of the four domains against the guidance in force now, before the questions arrive.