What Physics-Driven Peptide Design Does and Does Not Establish
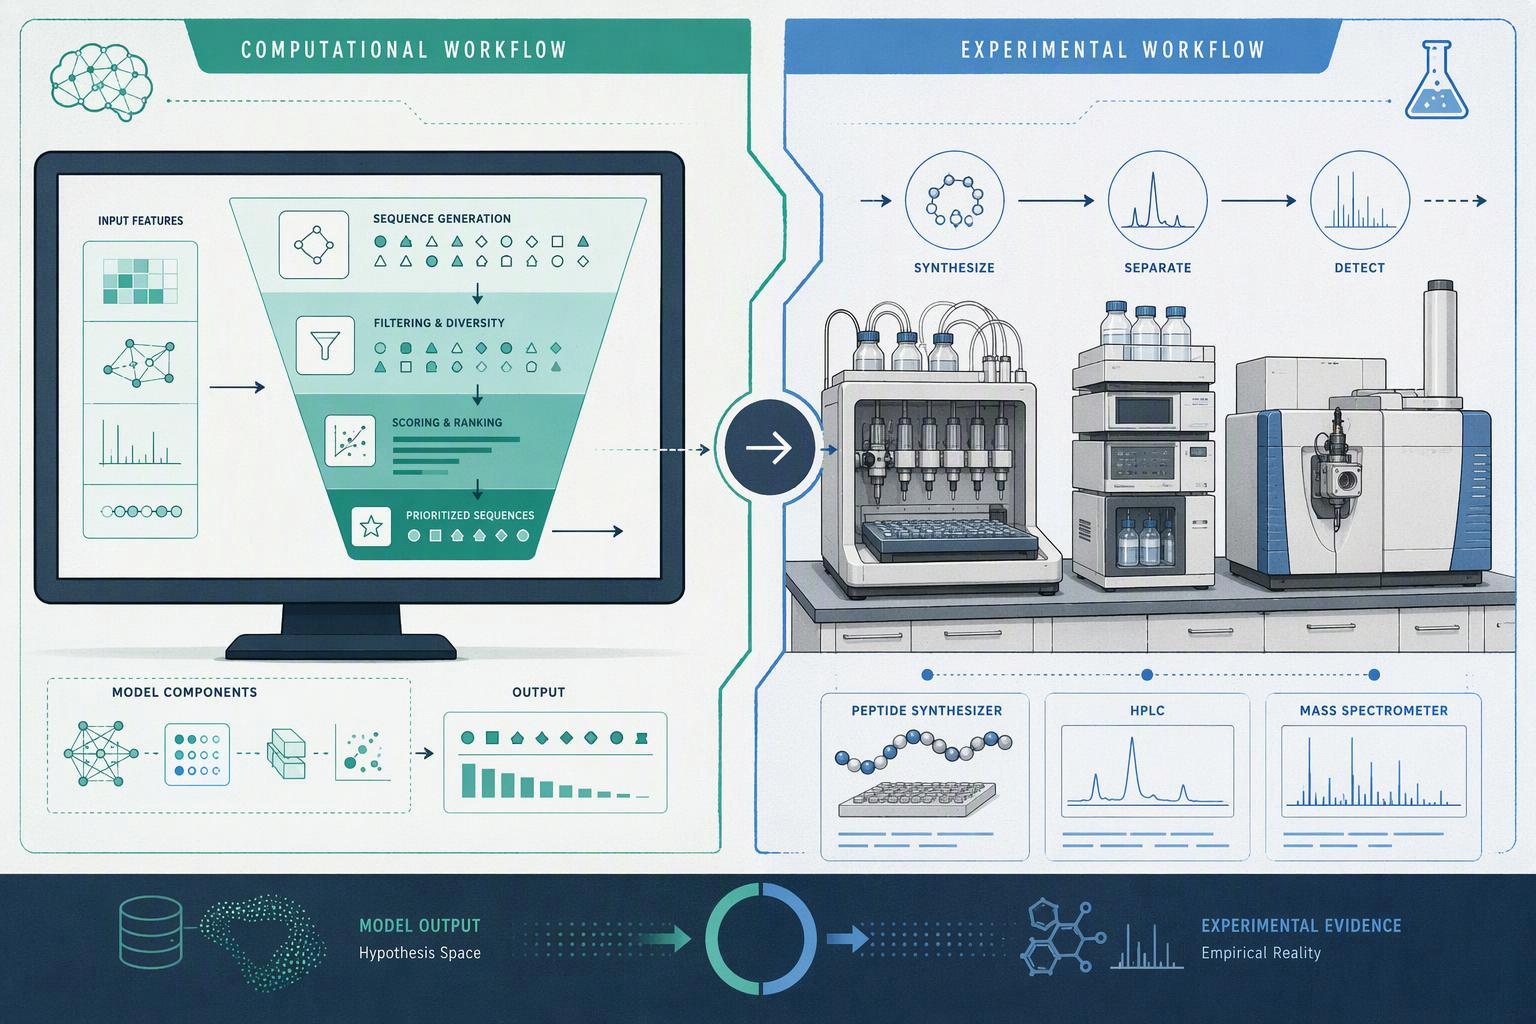
A computational score is a hypothesis about a modeled property. Validate it with the relevant experimental endpoint and separately assess synthesis feasibility, impurity risk and manufacturability. Do not transfer a published binder hit rate to a different peptide class or target without evidence.
Computational design success rates are comparable only when target, molecule class, selection process and experimental success criteria match. Older physics-based studies and newer binder pipelines use different settings; neither establishes a success rate for the construct being ordered.
Key Takeaway: A model score establishes plausibility, not identity. Only synthesis, purification and analytical confirmation establish that a molecule exists as specified.
Why Sequence Feasibility Comes Before Any Score
A peptide sequence feasibility assessment asks a different question from the one the design model answered. Predicted affinity says nothing about whether the chain will assemble on resin, and the two properties are largely independent: a high-scoring binder can be the hardest sequence in the set to make.
On-resin aggregation is the classic explanation for sequence-specific variability in synthetic efficiency, bisa lafazin Merck’s technical article on overcoming aggregation in SPPS (dawo da 2026-02-27). The same source identifies the recurring culprits: contiguous stretches of Ala, Val and Ile, and residues able to form intra-chain hydrogen bonds such as Gln, Ser and Thr. Ala-Gly recurs in hydrophobic transmembrane and amyloidogenic peptides, which is why designs drawn from membrane or aggregation-prone targets deserve a second look before they are quoted.
The failure is visible before the batch is lost. Severe aggregation shows as resin shrinkage in batch synthesis and a flattened or broadened deprotection profile in flow, and once the resin is collapsed, ninhydrin and TNBS coupling tests can return false negatives: the test reads as complete while the chain is truncated. For sequences longer than 20 amino acid, track assembly by small-scale TFA cleavage rather than trusting the colour test alone.
|
Screen |
What to record |
Readout that exposes a deletion or truncation |
|---|---|---|
|
Hydrophobic run length |
Longest contiguous Ala/Val/Ile stretch |
LC-MS mass short by one or more residues |
|
Aggregation-prone motifs |
Gln, Don zama, Thr clusters; Ala-Gly repeats |
Broadened or split HPLC main peak |
|
Chain length |
Total residues; position of the longest run |
Small-scale TFA cleavage profile at each segment |
|
Coupling performance |
Peptide Library Synthesis Deprotection profile shape in flow |
Flattened deprotection trace; false-negative Kaiser test |
The practical consequence for a design-to-synthesis handoff is that feasibility is a property of the sequence, not of its score. Screen the four rows Thermo Peptide Synthesis above before the sequence enters a quote, and flag any design that fails two or more of them for a route discussion rather than a straight order.
Modification Compatibility: What the Route Can Actually Carry
Peptide modification compatibility is a route question, not a catalogue question. Most non-canonical modifications leave the basic Fmoc-SPPS cycle intact, but each one adds a constraint the route has to absorb: stapling through Alloc/allyl or Mtt/Mmt/Dmt handles installed during assembly and reacted on-resin or late-stage, head-to-tail cyclization needing an orthogonal C-terminal anchor or a cleavable linker, D-amino acids needing milder coupling for racemization control, and N-methylated residues needing stronger activation, higher equivalents or double coupling.
Asp-containing motifs can be vulnerable to aspartimide formation during base exposure, with severity depending on sequence, protecting groups and deprotection conditions. Screen the actual construct and quantify relevant variants rather than using a per-cycle literature range as the process specification.
Screen every proposed modification against the route before you approve it: which handle it needs, whether that handle survives every subsequent deprotection, and what the final cleavage does to it.
Setting Purity Targets by Application, Not by Habit
Peptide purity targets by application should be set at order stage, because the grade you specify drives the synthesis route, the purification effort and the yield you can realistically recover. A standard vendor mapping such as GenScript’s recommended peptide purity guidelines places crude material at first-pass non-sensitive screening, >75% for ELISA, peptide arrays and polyclonal antibody antigen, >85% for in-vitro bioassays, epitope mapping and semi-quantitative enzyme-substrate work, >95% for quantitative receptor-ligand work, blocking or competitive inhibition, NMR and quantitative phosphorylation or proteolysis, kuma >98% for crystallography, GMP peptides, clinical trials and SAR. Crude peptides are typically around 50–60% pure, though that figure still needs batch verification before it goes into a publication.
|
Amfani da niyya |
Typical purity target |
|---|---|
|
First-pass, non-sensitive screening |
Danyen mai |
|
The Synthesis Of Aspirin ELISA, peptide arrays, polyclonal antibody antigen |
>75% |
|
In-vitro bioassays, epitope mapping, semi-quantitative enzyme-substrate work |
>85% |
|
Quantitative receptor-ligand work, blocking/competitive inhibition, NMR, quantitative phosphorylation or proteolysis Large Scale Peptide Synthesis |
>95% |
|
Crystallography, GMP peptides, clinical trials, SAR |
> Peptides ;98% |
Two cautions belong with that ladder. Na farko, purity is an HPLC area-percent of peptide-related material, while net peptide content is a separate quantity that accounts for counterion and water (Bachem, Quality Control of Amino Acids & Peptides, 2026). Na biyu, the vendor tables agree in band but are commercial sources with an interest in selling higher grades, so treat them as a starting framework rather than a specification.
Tabbatar da Nazari: Why One HPLC Number Is Not Identity
Peptide analytical confirmation by LC-MS starts from a hard limit: a single HPLC purity number cannot tell an isomeric modification apart from the intended one, because isomers and isobars share mass while differing in site or arrangement. HPLC measures chromatographic homogeneity, not exact chemical identity (Navigating Challenges in Mass Spectrometry Analysis of Endogenous and Synthetic Protein Modifications, PMC, 2026). A clean main peak that is not the intended molecule is a real failure mode, ba ka'ida ba.
Sample handling compounds the problem. Oxidation, deamidation, formylation and acetylation can arise during processing and mimic genuine modifications, confounding identification (same PMC review, 2026). Process control changes the size of that burden: for methionine-containing sequences, the oxidation side reaction fell from roughly 20% to about 5% under optimised conditions (Methionine-Containing Peptides: Avoiding Secondary Reactions, PMC, 2023). Peptide
Expect a defined impurity spectrum rather than a single contaminant: shafewa, tsinkewa, shigar, stereoisomer or racemization, cleavage-related and degradation products, plus high-molecular-weight species such as dimers and aggregates (Jagorar EMA akan haɓakawa da kera peptides na roba, 2022).
Don Tukwici: Pair a chromatographic purity figure with an orthogonal identity method before releasing a lot. One number describes how much material elutes together, not what that material is.
Bayanin iyaka: this article covers synthesis and analytical practice. It is not clinical or therapeutic guidance.
From Design Output to a Synthesis-Ready Specification
The physics-driven peptide design to synthesis handoff works when the specification, not the structure file, carries the design intent. A synthesis team should be able to start from the specification alone, without re-deriving why a residue or a modification was chosen.
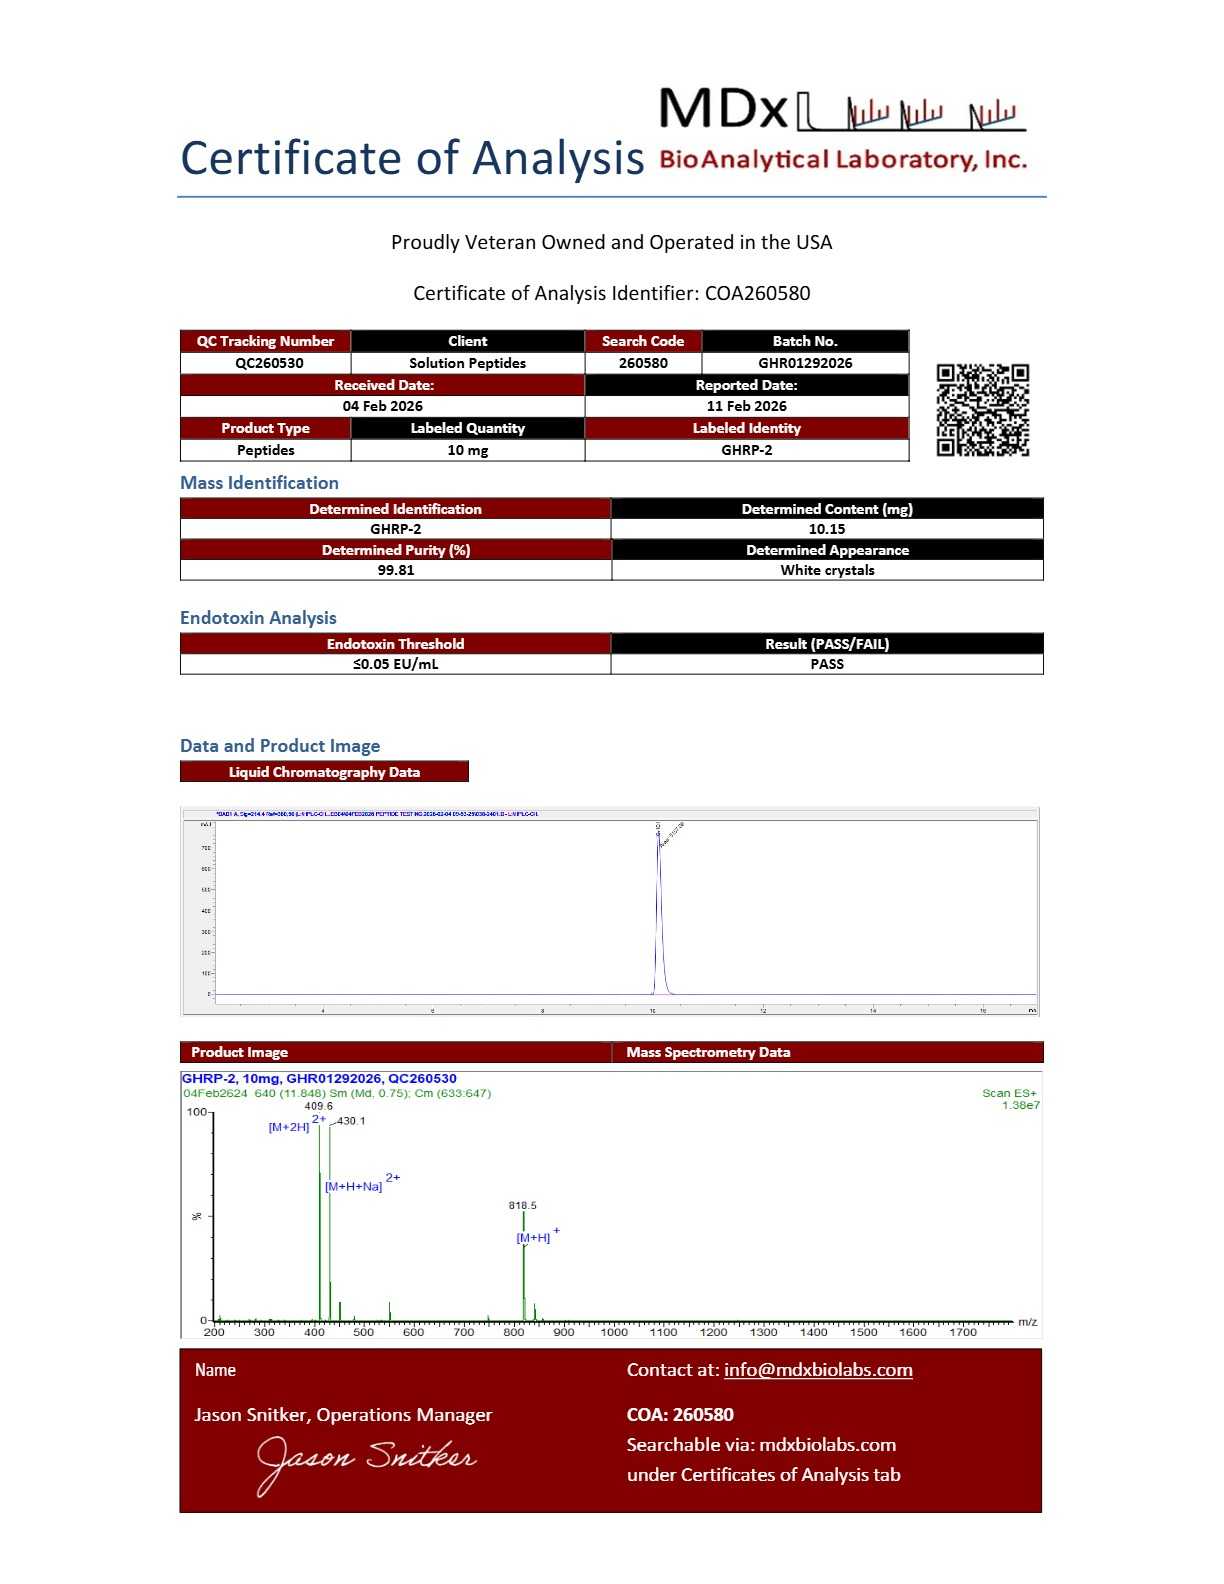
A peptide certificate of analysis typically carries HPLC purity as area percent, MS ainihi, net peptide content or amino acid analysis, sabani, abun ciki na ruwa, kuma, for higher grades, endotoxin, bioburden and residual solvent (Peptides na Amurka). Endotoxin limits are commonly specified by intended use rather than by a single fixed number: below 1 EU/mg for routine cell-based work, below 0.1 EU/mg for sensitive immune-cell assays, kuma 0.1 ku 0.5 EU/mg for preclinical in vivo material (Creative Proteomics).
Key Takeaway: The handoff package is the specification: sequence and modifications, route constraints, darajar tsarki, and the analytical package that will release the lot.
Jagorar EMA akan haɓakawa da kera peptides na roba, first published 9 Disamba 2025 and legally effective from 1 Yuni 2026, covers manufacturing process, halayyar, specifications and analytical control (EMA). An integrated platform such as MOL Changes shows the same contract in practice: sequence and intended use assessed at intake, then HPLC, MS and sterility testing reported with the lot.
Common Mistakes That Turn a Good Design into a Failed Batch
The most consequential mistake is treating an in silico score as experimental validation. A 2023 Nature Portfolio study of de novo designed protein interfaces reported 93.5% failure across heterodimeric interfaces and 96.4% across homodimeric interfaces (Communications Chemistry, 2023). That is one study, not a corroborated benchmark, but it makes the point: a favorable computational score predicts plausibility, not a synthesizable, kwayoyin halitta shirye-shirye.
The recurring bench mistakes follow the same pattern. Teams skip a feasibility screen on hydrophobic or aggregation-prone stretches, because the design output looked clean. They assume a modification is route-compatible without checking the protecting-group strategy it demands. They specify purity by habit rather than by application. And they accept a single HPLC purity number as identity confirmation, when purity and identity are separate questions.
Verify feasibility, route compatibility and the analytical acceptance criteria before committing a batch. Estimate rework time and cost from the actual sequence and supplier proposal; no universal resynthesis duration follows from these risks.
Tambayoyin da ake yawan yi
Can a physics-driven design score be used as evidence of activity?
A'a. A score from a physics-driven design tool is a hypothesis about structure, not a measurement of function. BindCraft’s own reported success rates for de novo binder design range from roughly 10% ku 100% depending on the target and the campaign (BindCraft), which is precisely why the score cannot stand in for an assay. Treat the computational result as a reason to synthesize and test, never as a substitute for the test.
How long does a peptide sequence feasibility assessment take, and what drives it?
Most of the time goes to the sequence itself rather than to queue position. Difficult stretches, aggregation-prone regions, and unusual modifications each add review time, because the assessment has to check coupling efficiency and route viability before anything is committed. A straightforward sequence with a standard purity target moves quickly; a heavily modified or long construct takes longer, and the honest answer is that the timeline follows the chemistry, not a fixed calendar.
Can a heavily modified or stapled construct be made by standard Fmoc SPPS?
Sau da yawa eh, but not automatically. Staples, macrocyclization, and unnatural residues each impose their own constraints on coupling and deprotection, and the route has to be chosen per sequence rather than assumed. This is exactly the kind of question a feasibility review exists to answer: whether the modification is compatible with standard Fmoc chemistry, or whether the route needs to change.
What should I do when the main peak does not match the expected mass?
Do not accept the peak on the basis of retention time alone. A mismatched mass means the identity is unconfirmed, and the correct response is to hold the lot and investigate rather than proceed. Purity by HPLC and identity by mass spectrometry answer different questions, and a clean chromatogram does not resolve a mass discrepancy.
Is a higher purity grade always worth the cost?
A'a. The right purity target follows the application, not a default preference for the highest number. A grade chosen for a demanding assay is money well spent; the same grade applied to a screening use may buy nothing the experiment needs. Set the target from what the downstream work actually requires.
Kammalawa
You can now screen a physics-driven peptide design to synthesis handoff in four moves: check sequence feasibility before you trust any score, confirm the route can carry each modification, set purity from the application rather than from habit, and require orthogonal analytical confirmation instead of a single HPLC number. The mechanism underneath all four is the same. A physics-driven design output is a hypothesis; kira, purification and analytical confirmation are what turn it into evidence.
MOL Changes offers custom peptide synthesis and analytical services. This article is educational and describes general synthesis and analytical practice; it is not medical advice.