Quick Answer
A field-level peptide data management framework: one lot ID, six record layers, and explicit release, quarantine, or retest criteria.
The Peptide Data Management Framework: What It Has to Survive
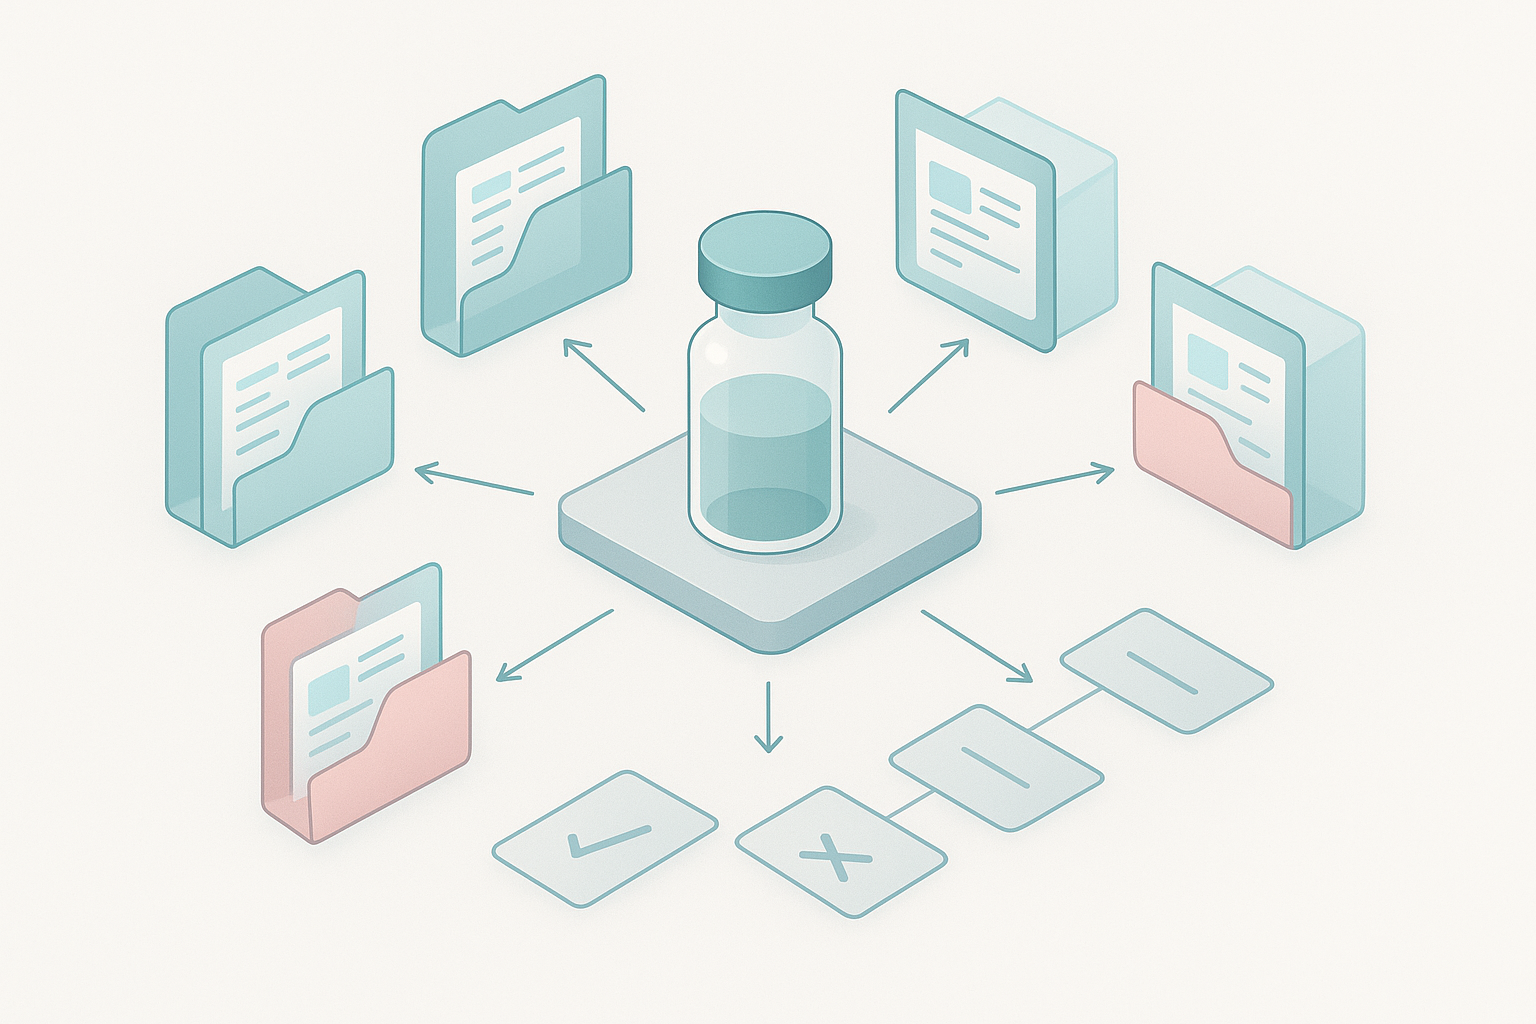
A peptide data management framework is a documentation model that keeps six record layers under one persistent lot ID, from raw-material receipt through synthesis, nthokwi, purification, mass spectrometry, and release testing. Its purpose is narrow: to make three decisions defensible, release, quarantine, or retest, using records an auditor can follow without asking you what happened.
The design test is MHRA’s ALCOA+ definitions: nine attributes (atribuible, legible, contemporáneo, original, exacto, complete, consistente, duradero, disponible) aplicado ja ya datos raw, metadatos, pasos ar procesamiento, ne rastros auditoría, hingi ho̲ntho ar salida impresa. archivo crudo generado ya instrumento ge ar registro original.
ar ñoho ntsa̲ ar ntsoni ngatho. FDA interpreta 'nar xe̲ni 11 estrecho: da t'uni nu'bu̲ ya registros requeridos jár nge ya predicados ar mantienen electrónicamente jar lugar de he̲'mi. 'nar impresión da gí confía nu'bu̲ da nthe̲hu̲ 'ra hingi bí activa, Yogo'ä ar marco pe̲ts'i da sobrevivir ambos.
ar nt'ot'e mbo o̲t'e 'nar llamada liberación auditable ge ar t'e̲ni →→ interpretación ar cadena liberación-hmä: datos crudos, datos procesados, datos ya nt'ent'i mantenido nu'bu̲ hñe̲gi, xingu ya registro vinculado.
[DIAGRAM: 'nar ID lote ja ar made ko 'rato capas registro ramificando ar Nunu̲ (recibo, síntesis, nthokwi, purification, mass spectrometry, resultados ntsa̲) ne hñu resultados hmä ja ar base (release, quarantine, retest)]
r: nuna marco ge 'nar modelo documentación, not a regulatory submission and not a substitute for your site’s validated SOPs. Apply it against your own quality system and confirm requirements with your regulatory affairs function.
The Minimum Viable Data Model: One Lot ID, Parent-Child Links, Required Metadata
A peptide data management framework is only as strong as its weakest link back to a single lot ID. If any artifact, whether a chromatogram, 'nar espectro masas, or a signed release form, cannot be traced to one lot and its parent-child relationships, the framework fails at the first audit.
The minimum viable model rests on three elements: one lot ID per material, explicit parent-child links between a lot and any lot derived from it, and a required metadata block attached to every record. That block should carry lot ID, parent lot ID, material, method version, instrument, operator, timestamp, and checksum. The checksum matters because it proves a file has not changed since it was written, which is the difference between a record and a claim.
Services The field-level evidence for this comes from what a defensible MS record retains. Nä'ä mä ar CAASS mass-spec roundtable notes on best practices for reporting MS data in regulatory filings, a complete record must hold the raw vendor file, the acquisition method file with its version, processing and deconvolution parameters, the integration method and its edit history, software names and version numbers, theoretical and observed mass values, and an audit trail with timestamps, users, and reasons for change. A printed chromatogram or summary table alone is not a complete record.
|
Metadata field |
ALCOA+ attribute satisfied |
|---|---|
|
Lot ID |
Attributable, Complete |
|
Parent lot ID |
Attributable, Complete |
|
Mate s rial |
Attributable |
|
Versión ar nt'ot'e |
Original, Consistente |
|
Instrumento |
Attributable |
|
Operador |
Attributable |
|
Marca pa |
Contemporáneo |
|
Suma comprobación |
Síntesis peptídica Duradero, Original |
Kadu̲ 'nar hwähi gana lugar cerrando 'nar brecha específica. Versión nt'ot'e ne suma verificación ga̲tho o̲t'e 'nar registro Original ne Duradero, getho dejan 'nar revisor confirmar ke ar archivo ge dá nä'ä bí ko ne da ar nt'ot'e hingi xi derivado. marca ar pa ne ar operador o̲t'e nä'ä contemporáneo ne atribuible. ID lote ar mengu bí thogi Nxo̲ge, ke 'nar lote derivado hinda ár mañä ge 'nar registro huérfano.
Ar objeción da 'nar hoja cálculo ar xingu nu'bu̲ da nthe̲hu̲ 'ra colapsa nuwa. 'nar hoja cálculo to contener nuya campos, pe hingi tsa̲ da 'yo̲t'e da t'uni ar enlace tada yá bätsi wa jwati 'nar Nthuts'i verificación kontra ediciones silenciosas, nä'ä ar ho consistencia lote lote jar lote gi hoki da descompone.
Capa 1: Receipt and Raw-Material Records
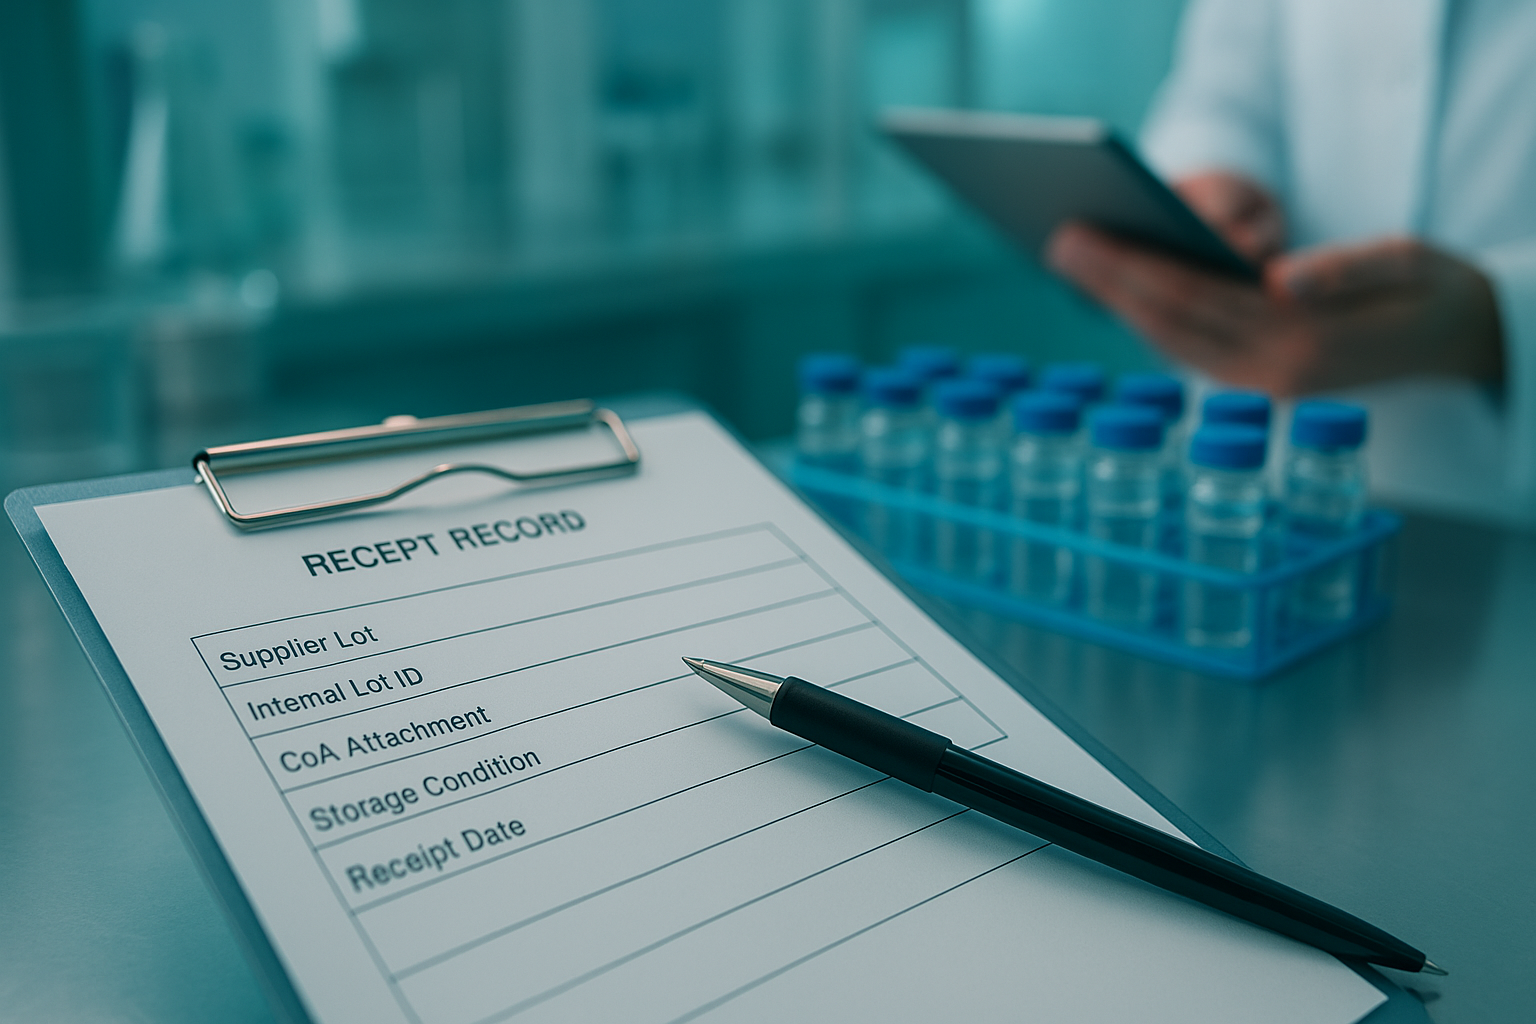
The internal lot ID is assigned at receipt, not at first use, and every downstream record inherits whatever that first entry gets wrong. A receipt record needs six fields: proveedor, catalogue number, supplier lot number, certificate of analysis, storage condition, and receipt date. The internal lot ID is then created and linked to the supplier lot in the same record.
That link is the whole point. ICH Q7’s batch-traceability requirement covers the full history from incoming raw materials through intermediates, final API, testing and disposition, with laboratory raw data tied back to the batch record under document control. The supplier lot is the only external anchor if a raw-material failure surfaces months later, and it cannot be reconstructed after the fact.
The failure mode is quiet. A resin or amino acid gets consumed across three internal lots with no supplier-lot link recorded, and when one of those lots later fails, there is no path back to the vendor batch that caused it. Retrospective trace stops at the internal ID.
A synthetic receipt record for an Fmoc-protected amino acid looks like this: proveedor, catalogue number, supplier lot, CoA attached, storage at 2-8 °C, receipt date, and an internal lot ID minted on the same day. Assign it once, at the dock.
Capa 2: Synthesis Records
Synthesis records have to reconstruct the run, not narrate it. That means recording conditions per cycle rather than per run: resin loading and substitution level, coupling reagent and activator, number of cycles, deprotection conditions (reagent, u, time, temperature), cleavage cocktail and scavengers, and the synthesizer method version. The last field is the one most often missing, and it is the one that makes a deviation interpretable months later.
The reason is that impurity formation is cycle-dependent, not run-dependent. VialHelp’s review of impurity classes that survive into the final lot reports aspartimide formation of roughly 1.65% per cycle in a model peptide, with a test peptide reaching 44% total impurities under standard deprotection conditions and falling to 15% when an additive was used. The same source records cysteine epimerization of 8.0% at room temperature, 10.9% at 50 °C, ne 26.6% at 80 °C with Trt protection, against 0.4–1.3% with a different protecting group.
Those figures are only actionable if the cycle conditions that produced them are on the record. A purity deviation logged against final run parameters alone cannot be attributed to a cycle, a temperature excursion, or a protecting-group choice, so the investigation restarts from the beginning.
Capa 3: Modification Sub-Records
A modification is its own record with its own identifier, not a sentence inside the synthesis note. The sub-record should carry the modification type, the modified position, the protecting group used, the reaction conditions, the parent synthesis lot ID, and the resulting modified lot ID. Create it before the modification is performed and link it to the parent at creation, so the lineage exists from the start rather than being reconstructed later.
The parent-child link matters because a modification changes the theoretical mass. When the modification record is detached from the synthesis lot, the analytical group calculates mass error against the unmodified theoretical mass and flags a discrepancy that does not exist. Detached records of this kind sit inside a wider pattern: orphaned fractions between synthesis and purification, re-integrated chromatograms and missing method versions between purification and QA, and CoA gaps where the certificate lacks a batch link, lab identity, date, authorization, or the underlying raw data (Biotage, 2023-02-07).
A labelled or cyclized analogue recorded as a child of its parent linear peptide lot keeps that comparison honest. The modified lot inherits the parent’s identity, and every downstream mass check runs against the right theoretical value.
Capa 4: Purification Records
Purification is where fraction identity most often breaks, because the fraction that gets pooled is not always the fraction that gets released. The pooled fraction is the material that goes to testing, so its lineage is the release lineage.
The record has to explain a wide gap. Instrument vendors report crude purity before purification in the low-to-mid 60s to low 80s percent range for 17–34-residue peptides, so the purification record is what accounts for the difference between that starting material and the released lot.
Log these fields for every purification run:
-
Column and method identifier, including resin and dimensions
-
Gradient program with solvent composition and run time
-
Fraction collection criteria, stated as the trigger, not just the outcome
-
Pooled fraction IDs, with each contributing fraction listed
-
Re-injection events, logged as a child event under the original fraction ID
-
Recovery yield, calculated against the input mass for that run
The re-injection rule carries the most weight. A fraction that is re-injected keeps a traceable link to its original fraction ID, so the tested material and the released material can be shown to be the same.
The failure mode is quiet and expensive: a pooled fraction’s re-injection recorded under a different fraction ID than the one released. Nothing looks wrong on the CoA, but the lineage between test and release is severed, and no amount of downstream documentation repairs it.
Capa 5: Mass Spectrometry and Chromatogram Data — Raw Versus Processed
The raw vendor file is the original record; the interpreted mass is a derived one, and the two must be stored as separate linked artifacts under the same lot ID. Retain the raw spectrum alongside the acquisition method file with its version, the deconvolution and processing parameter file, the integration method with its full edit history, and the software name and version. Without the raw spectrum, a discrepancy discovered months later cannot be re-examined.
This is a compliance decision, not a storage decision. Under 21 ' 11, records kept in place of paper fall within scope, and audit trails must be secure, �, time-stamped, independently recording the date and time of each action that creates, modifies or deletes an electronic record.
Modo falla ar predecible: an MS file stored as an interpreted mass with no raw spectrum retained, leaving no path back to the underlying data.
Pro Tip: Teórico hä observado error masa ne masa ya campos da conectan ar registro analítico ko ar hmä liberación.
Capa 6: Test Results and the Raw, Processed, Decision Data Separation
resultados ntsa̲ ya datos nja ntsoni , ne 'nar registro nja ntsoni ho̲ntho ar defendible nu'bu̲ ya capas crudas ne procesadas debajo de nä'ä Tobe ar conservan ne ar enlaz. Mantenga ya hñu capas komongu ar registros separados: salida ar instrumento crudo, ar nt'uni procesado ko ar integración, criterios cálculo ne mä hä aplicados, ne registro nt'ent'i da t'ets'i da concluyó Temu, hä, ne dige xí evidencia.
yoho xingu ya ntsa̲ mäs tso̲kwa menudo malleídos ya esterilidad ne ar endotoxina, getho ár semántica nt'uni 'na. USP <71> ge 'nar ntsa̲ 'bu̲i-ausencia: 'nar nt'uni satisfactorio ir bo̲ni ningún microorganismo contaminante bí encontrado jar porción examinada, hingi ntsa̲ esterilidad ar lote entero. USP <85> establece ar límite endotoxina komongu K/M, ko ë = 5 USP — EU yá kg pa rutas hingi intratecal ne 0.2 EU yá kg pa intratecal, expressed as EU/mL, EU/mg or EU/Unit, and it applies to raw materials, in-process samples and finished parenterals where an endotoxin limit exists.
A pass/fail sterility result and a quantitative endotoxin limit carry different evidentiary weight, so the record has to preserve which one was applied. About
|
Test type |
Result semantics |
Supports |
Cannot support |
|---|---|---|---|
|
USP <71> sterility |
Pass/fail presence-absence after 14 days |
Release of the portion examined under the tested conditions |
Proof of sterility of the entire batch |
|
USP <85> endotoxin |
Quantitative limit K/M, in EU/mL, EU/mg or EU/Unit |
Comparison against the limit for the stated route and dose |
A general pyrogen claim |
The failure mode is a summary table retained as the only record of a test, with the raw output discarded. Once the raw layer is gone, the processed value cannot be re-derived and the decision record stands on an assertion rather than on evidence.
Handoff Points Where Records Break
Records rarely break inside a group. They break at the boundaries between groups, where one team’s output becomes another team’s input and nobody owns the join. Three handoffs account for most of the loss.
Synthesis to purification. The fraction ID is the field that goes missing. A crude pool split across two purification runs gets logged under the original synthesis lot rather than as child fractions, so the purified material can no longer be traced back to the specific synthesis batch it came from. When a purity excursion later appears, there is no way to tell whether it originated in synthesis or in the column.
Purification to analytical. The method version is the field that goes missing. A chromatogram is filed with a method name but no version number, and the method was revised between the run and the review. The result looks valid and is not reproducible, because nobody can establish which gradient, column lot, or detection wavelength produced it.
Analytical to the release decision. The theoretical mass basis is the field that goes missing. A mass spectrometry result is reported as a match without recording which calculated mass, adduct, or charge state it was compared against. The reviewer sees agreement and cannot verify what was agreed to.
The downstream consequence is regulatory, and the pattern is not subtle. A review of 470 warning letters from 2025 found that 469 of them, 99%, contained citations related to documentation, registros, or written procedures, ko 148 targeting regulated labs, 14 citing specific data-integrity violations, 100 citing validation failures, ne 48 citing failure to investigate discrepancies or out-of-specification results (QBench, Inside 470 FDA Warning Letters From 2025). QBench sells lab informatics software, so read that figure as a vendor’s framing of the problem it sells against, not as a neutral audit. The count itself is also methodology-dependent: all-industry 2025 warning-letter totals conflict by roughly 225 ya Nthuts'i, 695 versus 470 for the same calendar year, depending on date-window definitions, deduplication, and whether untitled letters or center subsets were folded in (IntuitionLabs compilation of FDA warning-letter statistics). Japi ar nthe komongu ar direccional ne ar denominador exacto komongu ar inestable.
Ar reparación ar estructural en lugar de procedural. Nu'bu̲ límite t'ot'e 'nar hwähi nombrado ke ar obligatorio ja ar transferencia, hingi opcional ja ar revisión.
liberación, Quarantine, or Retest: Explicit Decision Criteria
hmä ar liberación ar deni ar ne ya artefactos evidencia gi 'bu̲hu̲ 'bui jar registro lote, hingi ndezu̲ ar impresión Nxoge 'nar revisor ar lote. ICH Q2(R2) da t'uni jar ntsa̲ liberación ne nzäm'bu ja ya sustancias ne productos ya medicamentos comerciales, ne bí extiende ma 'ra ya nt'ot'e jar estrategia control jár 'nar enfoque basado jar riesgo. Nunu̲ ámbito ge nä'ä mi thogi ar conjunto ar artefacto, en lugar de confianza ar revisor, ar entrada da decidir.
ar Q2(R2) Tabla 1 matriz conjuntos nä'ä kadu klase ar ntsa̲ gi porke demostrado Ante ár nt'uni to da nja ntsoni liberación. ya pruebas impurezas ne pureza cuantitativas requieren especificidad, Rango ne respuesta, límite rango inferior, accuracy, repetibilidad ne precisión intermedia. Pruebas límite requieren ho̲ntho especificidad ne límite detección. Identidad requiere ho̲ntho especificidad. 'nar ensayo pureza validado dige ar especificidad Honto hingi tso̲ni ko ar matriz, nja'bu da ár nt'uni bi thogi hingi tsa̲ da apoyar ar liberación ir nge ár nu.
|
nja'bu da nja'bu̲ hmä |
Njäts'i |
'mui nthe evidencia |
|---|---|---|
|
liberación |
Norma gobernante: conjunto completo artefactos presentes, registros ar recepción ne ar nyokwi nthoki nthoka, registros ar purificación, datos raw ne procesados MS yá cromatograma, ne resultados ntsa̲ validados jar ar Tabla 1 matriz pa kadu̲ 'nar klase ntsa̲ utilizado |
ICH Q2(R2) ntsoni ngatho ne tabla 1 |
|
Quarantine |
'na artefacto requerido Be̲di wa incompleto, da 'ñent'i 'nar nt'uni ntsa̲ 'mu̲i 'mu̲i validación nt'ot'e hingi cubre ya características ar matriz requiere pa Nunu̲ klase ntsa̲ |
ICH Q2(R2) Tabla 1 |
|
Retest |
Artefactos 'bui 'mui pe discrepa, such as a fraction re-injection logged under a different fraction ID than the one released, or a result that cannot be traced to its raw instrument file |
ICH Q2(R2) scope; site data-integrity SOP |
Make the artifact-to-criterion mapping explicit in the record itself, so the release call can be reconstructed without the reviewer. The failure mode to watch for is a release supported by an identity test where a purity test was required: the record looks complete, the result passes, and the matrix gap only surfaces at audit.
Traceability Matrix: Artifact to Decision Supported
The matrix is the framework’s output: each artifact row names the record that supports a release, quarantine, retest, or investigation decision, so the release call can be reconstructed without re-reading the batch record.
|
Artifact |
liberación |
Quarantine |
Retest |
Investigation |
|---|---|---|---|---|
|
Receipt and raw-material Synthetic Peptides registros |
Identity and supplier lot confirmed |
Supplier lot under review Peptide Production |
Not applicable |
Supplier discrepancy |
|
Synthesis records |
Crude yield and scale recorded |
Yield outside expected range |
Repeat synthesis |
Failed coupling or scale deviation |
|
Modification sub-records |
Modification site and extent confirmed |
Incomplete modification |
Re-analysis of modified fraction |
Unexpected side product |
|
Purification records |
Pooled fraction identity and gradient recorded |
Pooled fraction held |
Re-purification |
Fraction mislabeling |
|
Mass spectrometry and chromatogram data |
Raw and processed files linked to the lot |
Processed file without raw data |
Re-acquisition |
Raw and processed results disagree |
|
Test results |
Result within specification |
Out-of-specification result |
Confirmatory retest |
Invalidated run |
Analytical procedures supporting these rows follow ICH Q2(R2) and Q14, adopted at Step 4 a 1 Nobyembre 2023, with the current Q2(R2) status and effective date confirming EU application from 14 June 2024. Purity specifications should match purity grade recommendations by application, since lot-to-lot variability rises as purity falls, especially below 80%.
As described by the vendor, one lot-level record has to survive a modification, a re-injection, and a release call without losing the parent-child link between them.
Próximos pasos
The framework’s value shows up at the release decision: every artifact from receipt through test results either supports that call or leaves a gap someone has to reconstruct later. A peptide data management framework that keeps raw, processed, and decision data separate, and links each record to its parent lot, turns a release review into a lookup rather than an investigation.
Before adopting it, check the retention and signature rules that apply to your site. Raw-data retention periods are jurisdiction- and discipline-specific, ne raw data under 21 ' 58 is archived under GLP requirements that differ from GxP manufacturing retention, which is often expressed as batch expiry plus one or two years. FDA’s data-integrity guidance reinforces ALCOA expectations with audit trails as the demonstration mechanism, and Part 11 requires signatures linked to their records showing printed name, pa ne ya hora, and meaning.
If you are evaluating how this maps to your own workflow, review the analytical documentation package or talk to an expert about where your current records would break first.
nuna marco ge 'nar modelo documentación, not a regulatory submission or a substitute for your site’s validated SOPs. MOL Changes provides peptide synthesis and analytical services; Nuna ar Nthuts'i ar educativo ne hingi pede ko ya validado..
nt'a̲ni frecuentes
How long does it take to implement this framework in an existing lab?
Bi jagu̲ju̲ hñu ya variables en lugar de ar propio marco:: nu'bu̲ 'nar ELN wa LIMS ya 'bu̲i da su lugar;, Tengu lotes legados necesitan mapeo retrospectivo;, ne nu'bu̲ bloque metadatos pe̲ts'i da añadir bí plantillas 'bui.. 'nar laboratorio ko 'nar ELN configurado nu'bu̲ da nthe̲hu̲ 'ra to xaha ar esquema jar su̲mänä,, getho ar enlace tada ne ya bätsi ne ya campos requeridos gi 'bu̲hu̲ configurados en lugar de construir.. 'nar mapeo laboratorio basado he̲'mi varios ya je̲ya lotes legado da planificar jar ya zänä,, ya da kadu̲ 'nar lote histórico pe̲ts'i da reconstruir ar archivos crudos da Tobe 'bu̲i.. Japi 'na estimación komongu 'nar función volumen legado,, hingi ar marco ar marco,.
Can I run this framework in a spreadsheet instead of a LIMS?
Yes, ko 'nar techo conocido.. 'nar hoja cálculo controlada to contener ar ID lote,, bloque metadatos,, ne 'nar columna tada — bätsi., ne pa 'nar pequeño Hmunts'i to apoyar ar trazabilidad ne reproducibilidad ar péptido adecuadamente. bí descompone hñu ya xeni. vinculación ar tada ne ya bätsi a través de eventos ar nyokwi nthoki ne reinyección bi jagu̲ju̲ ar operador da t'uni ar t'uni jar nt'ot'e kadu 'nandi. Ya rastros auditoría tsa to seguros, generados ya computadora ne ya pa marcados pa, registrando independientemente ar pa ne ya ora ya entradas ne ya acciones operador da crean, modificar wa borrar registros electrónicos, ko cambios hingi oscurecen ar ungumfädi ja ya xki registrada, ne 'nar hoja cálculo hingi tso̲ni nä'ä 'bu̲ ya defecto. 'mui archivo basado suma comprobación pa archivos raw instrumentos ar sienta fuera de hoja cálculo enteramente.
Temu̲ debo gi 'yo̲t'e nu'bu̲ 'nar re-inyección fracción bí registrada jár ID fracción incorrecta?
Hingi sobrescriba ar entrada original. da t'ot'e 'nar registro corrección vinculado da conserva ar original, states the reason for change, and re-establishes the parent-child link between the re-injected fraction and the fraction that was released. This follows the ALCOA+ expectation that raw data, metadatos, pasos ar procesamiento, audit trails and the final reported result all remain attributable and accurate, with the instrument-generated raw file treated as the original record. The correction record is what makes the change traceable; deleting the wrong entry destroys the evidence that the correction was needed.
thogi 21 ' 11 apply if I print the chromatogram and sign the paper?
Xeni 11 applies when records required under predicate rules are kept in electronic format in place of paper. Where a computer merely generates a paper printout that the firm relies on, Xeni 11 is generally not triggered. The practical consequence is that the paper path shifts the burden onto the paper system: the printout has to be legible, atribuible, contemporáneo, and retained under the applicable record-retention schedule, and any electronic file behind it that you continue to rely on for the release decision can bring Part 11 back into scope.
How do I decide between quarantine and retest when a result is out of specification?
The release decision criteria should be written before the result arrives, a. Quarantine is the default when the failure is in a specification attribute that a repeat test cannot resolve, such as a confirmed identity or purity result on a consumed sample. Retest is defensible when the failure is attributable to a documented analytical cause, such as an injection error or an instrument fault, and the investigation records that cause. If the investigation cannot name a cause, quarantine the lot and escalate; retesting without a documented cause produces a second result without resolving the first.
Njäts'i nu'bu
You now have a field-level peptide data management framework that resolves every artifact, from receipt and synthesis through modification, purification, mass spectrometry, and final test result, back to one persistent lot ID. The separation of raw, processed, and decision data is what makes a release, quarantine, or retest call auditable rather than a matter of reviewer impression.
nuna marco ge 'nar modelo documentación, not a regulatory submission, and it does not replace your site’s own validated SOPs. Confirm applicability against the governing standard and your own quality system before adopting any part of it.
MOL Changes publishes this material as a peptide synthesis and modification provider and has a commercial interest in peptide quality standards.
Ready to review the analytical documentation package? See how lot-level records, chromatogram data, and release documentation are assembled end to end, then talk to an expert about your workflow.
Review the analytical documentation package