Il quadro di gestione dei dati sui peptidi: Cosa deve sopravvivere
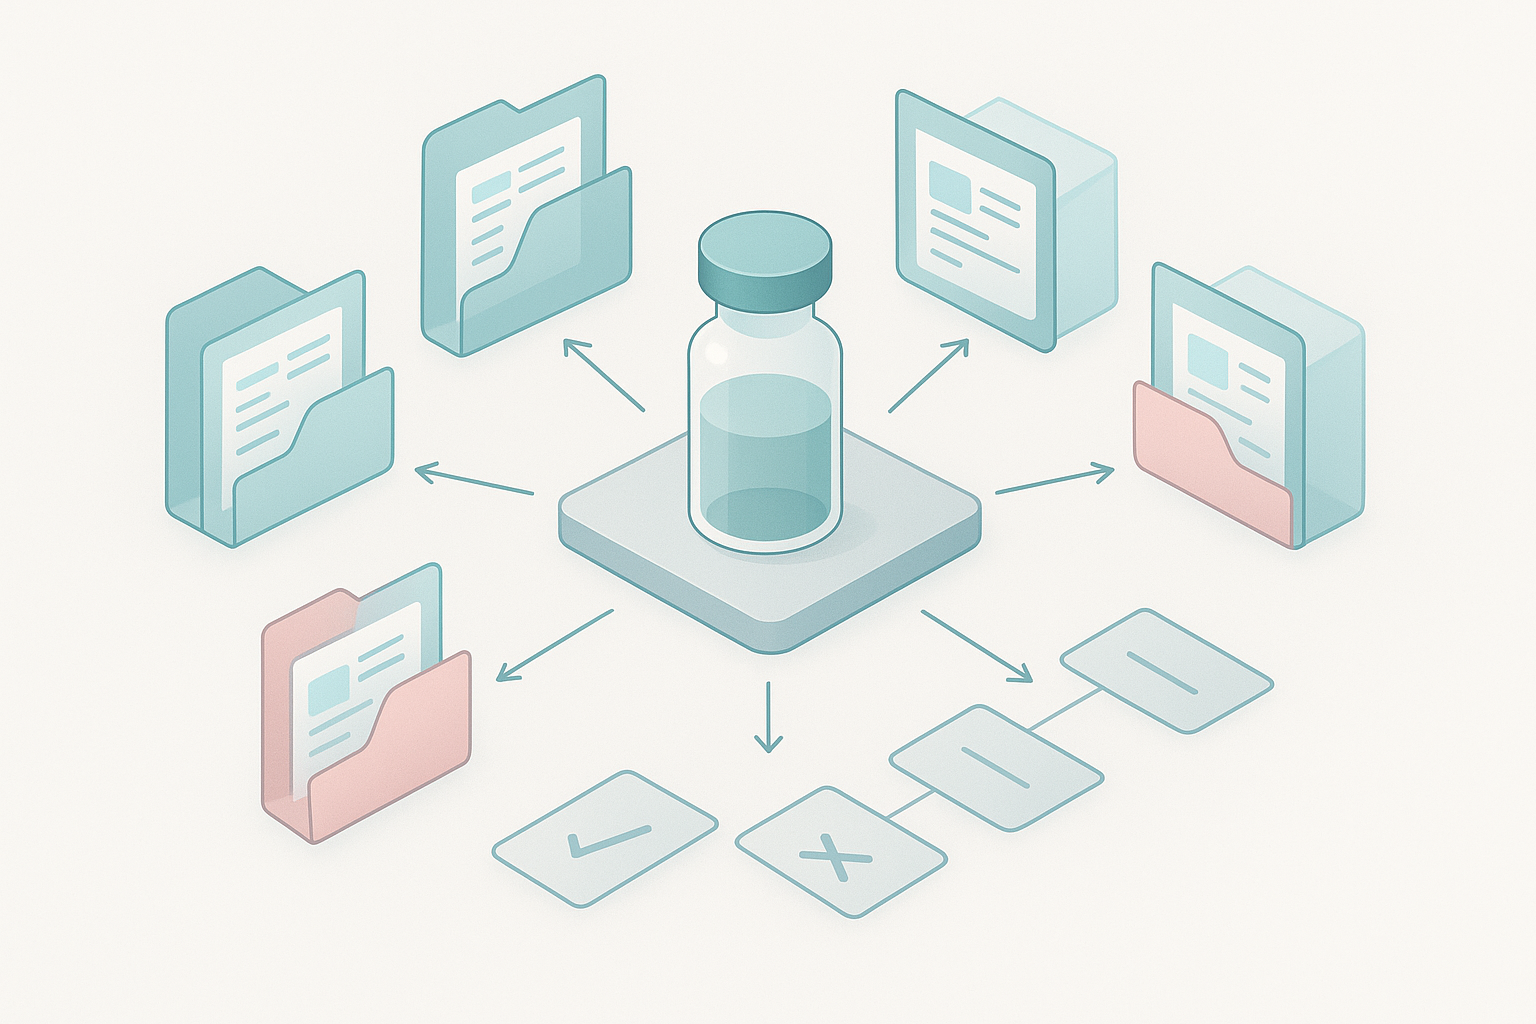
Un framework di gestione dei dati peptidici è un modello di documentazione che mantiene sei livelli di record sotto un unico ID lotto persistente, dal ricevimento delle materie prime alla sintesi, modifica, purificazione, spettrometria di massa, e test di rilascio. Il suo scopo è ristretto: rendere tre decisioni difendibili, pubblicazione, quarantena, o ripetere il test, utilizzando le registrazioni che un revisore può seguire senza chiederti cosa è successo.
Il test di progettazione è Definizioni ALCOA+ di MHRA: nove attributi (attribuibile, leggibile, contemporaneo, originale, accurato, completare, coerente, duraturo, disponibile) applicata ai dati grezzi, metadati, fasi di lavorazione, e piste di controllo, non solo l'output stampato. Il file raw generato dallo strumento è il record originale.
Il secondo test è la portata. La FDA interpreta la Parte 11 in modo restrittivo: si applica quando le registrazioni previste dalle norme presupposti sono conservate in formato elettronico anziché cartaceo. Una stampa su cui fai affidamento generalmente non la attiva, ed è per questo che la struttura deve sopravvivere ad entrambi.
Il meccanismo che rende verificabile una chiamata di rilascio è la catena misurazione → interpretazione → rilascio-decisione: dati grezzi, dati trattati, e i dati decisionali mantenuti separati, tipi di record collegati.
[DIAGRAMMA: Un ID lotto al centro da cui si diramano sei livelli di record (ricevuta, sintesi, modifica, purificazione, spettrometria di massa, risultati dei test) e tre esiti decisionali alla base (pubblicazione, quarantena, ripetere il test)]
⚠️ Attenzione: Questo framework è un modello di documentazione, non si tratta di una presentazione normativa e non di un sostituto delle SOP convalidate del tuo sito. Applicatelo al vostro sistema di qualità e confermate i requisiti con la vostra funzione affari regolatori.
Il modello minimo di dati vitali: Un ID lotto, Collegamenti genitore-figlio, Metadati obbligatori
Un framework di gestione dei dati peptidici è forte quanto il suo collegamento più debole a un singolo ID lotto. Se qualche artefatto, se un cromatogramma, uno spettro di massa, o una liberatoria firmata, non può essere ricondotto a un lotto e alle sue relazioni genitore-figlio, il quadro fallisce al primo audit.
Il modello minimo praticabile si basa su tre elementi: un ID lotto per materiale, collegamenti genitore-figlio espliciti tra un lotto e qualsiasi lotto da esso derivato, e un blocco di metadati obbligatorio allegato a ogni record. Quel blocco dovrebbe riportare l'ID del lotto, ID lotto principale, materiale, versione del metodo, strumento, operatore, timestamp, e somma di controllo. Il checksum è importante perché dimostra che un file non è cambiato da quando è stato scritto, che è la differenza tra un record e un reclamo.
Servizi La prova sul campo di ciò deriva da ciò che conserva un record difendibile della SM. Secondo il Note della tavola rotonda sulle specifiche di massa del CAASS sulle migliori pratiche per la segnalazione dei dati degli Stati membri nei documenti normativi, un record completo deve contenere il file grezzo del fornitore, il file del metodo di acquisizione con la sua versione, parametri di elaborazione e deconvoluzione, il metodo di integrazione e la cronologia delle modifiche, nomi del software e numeri di versione, valori di massa teorici e osservati, e una traccia di controllo con timestamp, utenti, e ragioni del cambiamento. Un cromatogramma stampato o una tabella riepilogativa da soli non costituiscono una registrazione completa.
|
Campo metadati |
Attributo ALCOA+ soddisfatto |
|---|---|
|
Identificativo del lotto |
Attribuibile, Completare |
|
ID lotto principale |
Attribuibile, Completare |
|
Compagno Negozio rial |
Attribuibile |
|
Versione del metodo |
Originale, Coerente |
|
Strumento |
Attribuibile |
|
Operatore |
Attribuibile |
|
Timestamp |
Contemporaneo |
|
Somma di controllo |
Sintesi peptidica Duraturo, Originale |
Ogni campo guadagna il suo posto colmando una lacuna specifica. La versione del metodo e il checksum insieme creano un record originale e duraturo, perché consentono a un revisore di confermare che il file è quello che è stato scritto e che il metodo non è andato alla deriva. Timestamp e operatore lo rendono contemporaneo e attribuibile. L'ID lotto principale lo rende completo, poiché un lotto derivato senza il relativo genitore è un record orfano.
L'obiezione secondo cui un foglio di calcolo è sufficiente di solito crolla qui. Un foglio di calcolo può contenere questi campi, ma non può imporre il collegamento genitore-figlio o preservare un checksum contro le modifiche silenziose, che è dove la coerenza tra batch tende a crollare.
Strato 1: Registrazioni delle ricevute e delle materie prime
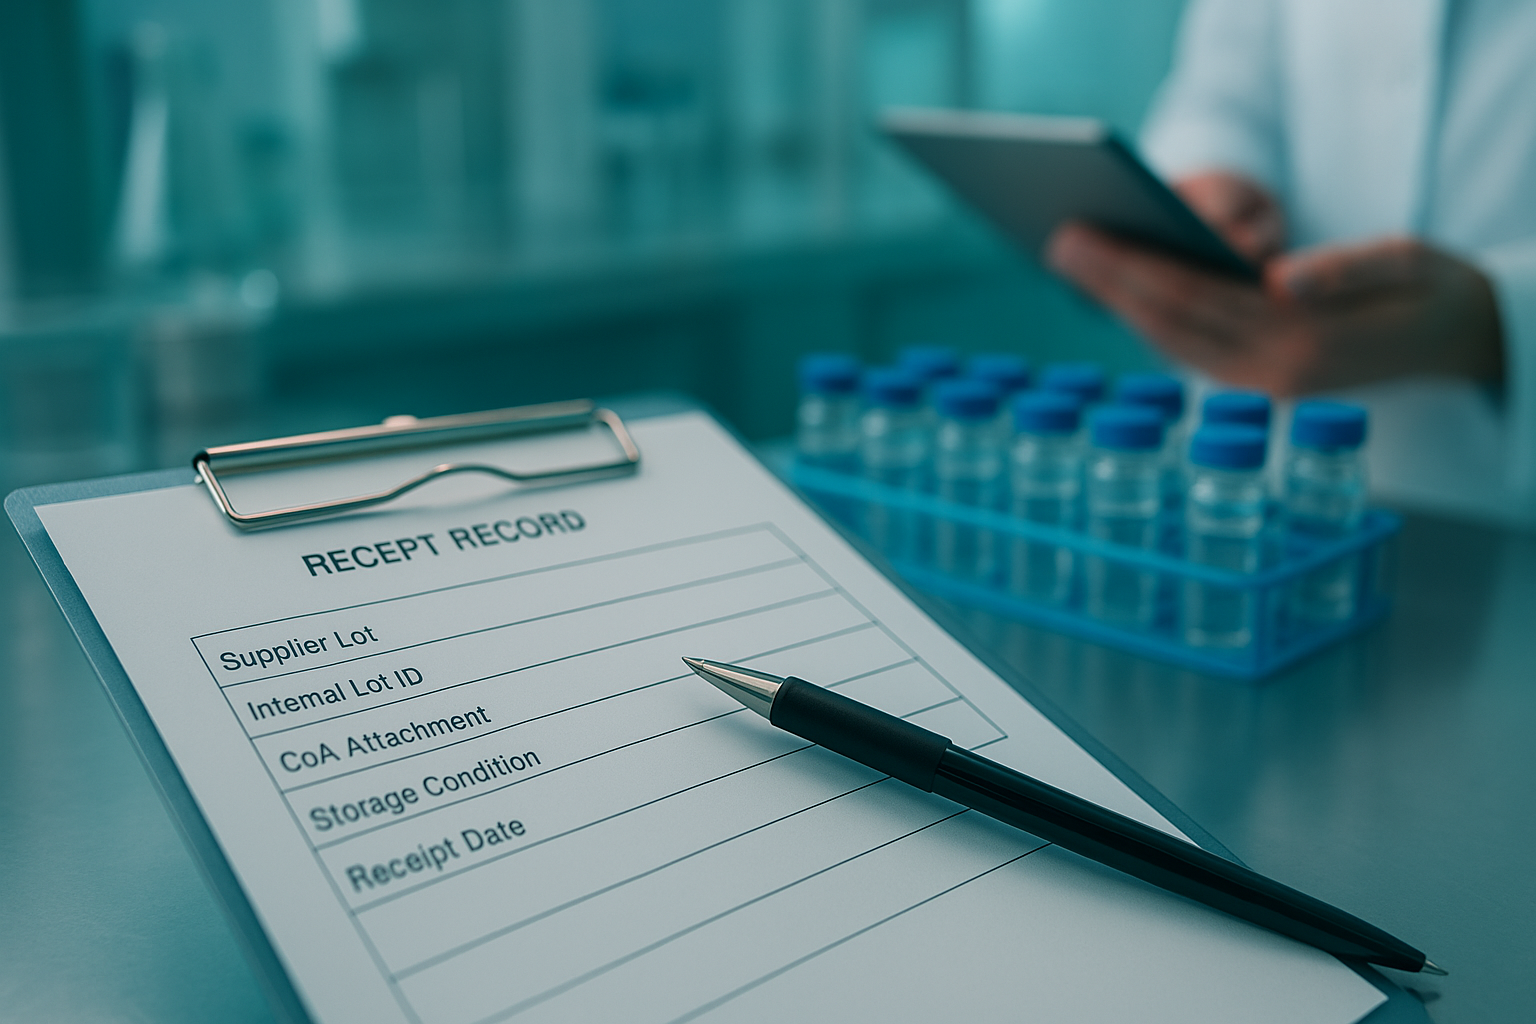
L'ID lotto interno viene assegnato al ricevimento, non al primo utilizzo, e ogni record downstream eredita tutto ciò che la prima voce sbaglia. Un record di ricevuta necessita di sei campi: fornitore, numero di catalogo, numero di lotto del fornitore, certificato di analisi, condizione di conservazione, e data di ricevuta. L'ID lotto interno viene quindi creato e collegato al lotto del fornitore nello stesso record.
Questo collegamento è il punto centrale. Requisito di tracciabilità dei lotti di ICH Q7 copre l'intera storia dalle materie prime in entrata fino agli intermedi, API finale, test e disposizione, con dati grezzi di laboratorio legati al record del lotto sotto controllo documentale. Il lotto del fornitore è l’unico ancoraggio esterno nel caso in cui un guasto della materia prima emerga mesi dopo, e non può essere ricostruito a posteriori.
La modalità di errore è silenziosa. Una resina o un amminoacido viene consumato in tre lotti interni senza che venga registrato alcun collegamento fornitore-lotto, e quando uno di quei lotti successivamente fallisce, non è possibile risalire al batch del fornitore che l'ha causato. La traccia retrospettiva si ferma all'ID interno.
Un record di ricevuta sintetica per un amminoacido protetto da Fmoc si presenta così: fornitore, numero di catalogo, lotto del fornitore, CoA allegato, deposito a 2-8 °C, data di ricevimento, e un ID lotto interno coniato lo stesso giorno. Assegnalo una volta, al molo.
Strato 2: Record di sintesi
I record di sintesi devono ricostruire la corsa, non raccontarlo. Ciò significa registrare le condizioni per ciclo anziché per corsa: livello di caricamento e sostituzione della resina, reagente di accoppiamento e attivatore, numero di cicli, condizioni di deprotezione (reagente, concentrazione, tempo, temperatura), cocktail di scissione e spazzini, e la versione del metodo del sintetizzatore. L'ultimo campo è quello che più spesso manca, ed è proprio lei a rendere interpretabile una deviazione a distanza di mesi.
Il motivo è che la formazione di impurità dipende dal ciclo, non dipendente dalla corsa. La recensione di VialHelp di classi di impurità che sopravvivono nel lotto finale riporta una formazione di aspartimide di circa 1.65% per ciclo in un peptide modello, con un peptide di prova che raggiunge 44% impurità totali in condizioni di deprotezione standard e cadenti 15% quando è stato utilizzato un additivo. La stessa fonte registra l'epimerizzazione della cisteina 8.0% a temperatura ambiente, 10.9% A 50 °C, E 26.6% A 80 °C con protezione Trt, contro lo 0,4–1,3% con un diverso gruppo protettivo.
Tali cifre sono utilizzabili solo se le condizioni del ciclo che le hanno prodotte sono documentate. Una deviazione della purezza registrata rispetto ai soli parametri della corsa finale non può essere attribuita a un ciclo, un'escursione termica, o una scelta del gruppo protettore, quindi l'indagine ricomincia dall'inizio.
Strato 3: Sottorecord di modifica
Una modifica è il proprio record con il proprio identificatore, non una frase all'interno della nota di sintesi. Il sottorecord deve riportare il tipo di modifica, la posizione modificata, il gruppo protettivo utilizzato, le condizioni di reazione, l'ID del lotto della sintesi madre, e l'ID lotto modificato risultante. Crealo prima che venga eseguita la modifica e collegalo al genitore al momento della creazione, quindi il lignaggio esiste fin dall'inizio anziché essere ricostruito in seguito.
Il collegamento genitore-figlio è importante perché una modifica cambia la massa teorica. Quando il record di modifica viene staccato dal lotto di sintesi, il gruppo analitico calcola l'errore di massa rispetto alla massa teorica non modificata e segnala una discrepanza che non esiste. I record distaccati di questo tipo si collocano all'interno di uno schema più ampio: frazioni orfane tra sintesi e purificazione, cromatogrammi reintegrati e versioni di metodi mancanti tra purificazione e QA, e lacune CoA in cui il certificato non dispone di un collegamento batch, identità del laboratorio, data, autorizzazione, o i dati grezzi sottostanti (Biotage, 2023-02-07).
Un analogo marcato o ciclizzato registrato come figlio del lotto di peptidi lineari genitore mantiene onesto il confronto. Il lotto modificato eredita l’identità del genitore, e ogni controllo di massa a valle viene eseguito rispetto al giusto valore teorico.
Strato 4: Documenti di purificazione
La purificazione è il punto in cui l'identità della frazione più spesso si rompe, perché la frazione che viene messa in comune non è sempre la frazione che viene rilasciata. La frazione raggruppata è il materiale che viene sottoposto a test, quindi il suo lignaggio è il lignaggio del rilascio.
La documentazione deve spiegare un ampio divario. Instrument vendors report crude purity before purification in the low-to-mid 60s to low 80s percent range for 17–34-residue peptides, so the purification record is what accounts for the difference between that starting material and the released lot.
Log these fields for every purification run:
-
Column and method identifier, including resin and dimensions
-
Gradient program with solvent composition and run time
-
Fraction collection criteria, stated as the trigger, not just the outcome
-
Pooled fraction IDs, with each contributing fraction listed
-
Re-injection events, logged as a child event under the original fraction ID
-
Recovery yield, calculated against the input mass for that run
The re-injection rule carries the most weight. A fraction that is re-injected keeps a traceable link to its original fraction ID, so the tested material and the released material can be shown to be the same.
The failure mode is quiet and expensive: a pooled fraction’s re-injection recorded under a different fraction ID than the one released. Nothing looks wrong on the CoA, but the lineage between test and release is severed, and no amount of downstream documentation repairs it.
Strato 5: Dati della spettrometria di massa e del cromatogramma: grezzi e elaborati
The raw vendor file is the original record; the interpreted mass is a derived one, and the two must be stored as separate linked artifacts under the same lot ID. Retain the raw spectrum alongside the acquisition method file with its version, the deconvolution and processing parameter file, the integration method with its full edit history, and the software name and version. Without the raw spectrum, a discrepancy discovered months later cannot be re-examined.
This is a compliance decision, not a storage decision. Sotto 21 CFR parte 11, records kept in place of paper fall within scope, and audit trails must be secure, generato dal computer, time-stamped, independently recording the date and time of each action that creates, modifica o cancella un record elettronico.
The failure mode is predictable: an MS file stored as an interpreted mass with no raw spectrum retained, leaving no path back to the underlying data.
Per Suggerimento: Theoretical versus observed mass and mass error are the fields that connect the analytical record to the release decision.
Strato 6: Risultati dei test e dati grezzi, Elaborato, Separazione dei dati decisionali
Test results are decision data, and a decision record is only defensible when the raw and processed layers beneath it are still retained and linked. Keep the three layers as separate records: the raw instrument output, the processed result with integration, calculation and acceptance criteria applied, and the decision record naming who concluded what, Quando, and on which evidence.
The two test types most often misread are sterility and endotoxin, because their result semantics differ. USP <71> is a presence-absence test: a satisfactory result means no contaminating microorganism was found in the portion examined, not proof of sterility of the entire batch. USP <85> sets the endotoxin limit as K/M, with K = 5 USP-EU/kg for routes other than intrathecal and 0.2 EU/kg for intrathecal, expressed as EU/mL, EU/mg or EU/Unit, and it applies to raw materials, in-process samples and finished parenterals where an endotoxin limit exists.
A pass/fail sterility result and a quantitative endotoxin limit carry different evidentiary weight, so the record has to preserve which one was applied. Di
|
Test type |
Result semantics |
Supports |
Cannot support |
|---|---|---|---|
|
USP <71> sterilità |
Pass/fail presence-absence after 14 giorni |
Release of the portion examined under the tested conditions |
Proof of sterility of the entire batch |
|
USP <85> endotossina |
Quantitative limit K/M, in EU/mL, EU/mg or EU/Unit |
Comparison against the limit for the stated route and dose |
A general pyrogen claim |
The failure mode is a summary table retained as the only record of a test, with the raw output discarded. Once the raw layer is gone, the processed value cannot be re-derived and the decision record stands on an assertion rather than on evidence.
Punti di passaggio in cui i record vengono battuti
Records rarely break inside a group. They break at the boundaries between groups, where one team’s output becomes another team’s input and nobody owns the join. Three handoffs account for most of the loss.
Synthesis to purification. The fraction ID is the field that goes missing. A crude pool split across two purification runs gets logged under the original synthesis lot rather than as child fractions, so the purified material can no longer be traced back to the specific synthesis batch it came from. When a purity excursion later appears, there is no way to tell whether it originated in synthesis or in the column.
Purification to analytical. The method version is the field that goes missing. A chromatogram is filed with a method name but no version number, and the method was revised between the run and the review. The result looks valid and is not reproducible, because nobody can establish which gradient, column lot, or detection wavelength produced it.
Analytical to the release decision. The theoretical mass basis is the field that goes missing. A mass spectrometry result is reported as a match without recording which calculated mass, adduct, or charge state it was compared against. The reviewer sees agreement and cannot verify what was agreed to.
The downstream consequence is regulatory, and the pattern is not subtle. Una recensione di 470 warning letters from 2025 trovato quello 469 of them, 99%, contained citations related to documentation, records, or written procedures, con 148 targeting regulated labs, 14 citing specific data-integrity violations, 100 citing validation failures, E 48 citing failure to investigate discrepancies or out-of-specification results (QBench, Inside 470 FDA Warning Letters From 2025). QBench sells lab informatics software, so read that figure as a vendor’s framing of the problem it sells against, not as a neutral audit. The count itself is also methodology-dependent: all-industry 2025 warning-letter totals conflict by roughly 225 letters, 695 contro 470 for the same calendar year, depending on date-window definitions, deduplication, and whether untitled letters or center subsets were folded in (IntuitionLabs compilation of FDA warning-letter statistics). Treat the ratio as directional and the exact denominator as unsettled.
The fix is structural rather than procedural. Each boundary needs a named field that is mandatory at transfer, not optional at review.
Pubblicazione, Quarantena, o ripetere il test: Criteri decisionali espliciti
The release decision follows from which evidence artifacts are present in the lot record, not from a reviewer’s overall impression of the batch. Io Q2(R2) applies to release and stability testing of commercial drug substances and products, and it extends to other procedures in the control strategy under a risk-based approach. That scope is what makes the artifact set, rather than the reviewer’s confidence, the deciding input.
IL Q2(R2) Tavolo 1 matrix sets what each test type must have demonstrated before its result can carry a release decision. Quantitative impurity and purity tests require specificity, range and response, lower range limit, precisione, repeatability and intermediate precision. Limit tests require only specificity and detection limit. Identity requires only specificity. A purity assay validated on specificity alone does not meet the matrix, so its passing result cannot support release on its own.
|
Decision |
Evidence condition |
Governing standard |
|---|---|---|
|
Pubblicazione |
Full artifact set present: receipt and raw-material records, synthesis and modification records, purification records, raw and processed MS/chromatogram data, and test results validated to the Table 1 matrix for each test type used |
Io Q2(R2) scope and Table 1 |
|
Quarantena |
Any required artifact missing or incomplete, including a test result whose method validation does not cover the characteristics the matrix requires for that test type |
Io Q2(R2) Tavolo 1 |
|
Retest |
Artifacts present but discrepant, such as a fraction re-injection logged under a different fraction ID than the one released, or a result that cannot be traced to its raw instrument file |
Io Q2(R2) scope; site data-integrity SOP |
Make the artifact-to-criterion mapping explicit in the record itself, so the release call can be reconstructed without the reviewer. The failure mode to watch for is a release supported by an identity test where a purity test was required: the record looks complete, the result passes, and the matrix gap only surfaces at audit.
Matrice di tracciabilità: Artefatto per decisione supportata
The matrix is the framework’s output: each artifact row names the record that supports a release, quarantena, ripetere il test, or investigation decision, so the release call can be reconstructed without re-reading the batch record.
|
Artefatto |
Pubblicazione |
Quarantena |
Retest |
Investigation |
|---|---|---|---|---|
|
Receipt and raw-material Peptidi sintetici records |
Identity and supplier lot confirmed |
Supplier lot under review Produzione di peptidi |
Not applicable |
Supplier discrepancy |
|
Synthesis records |
Crude yield and scale recorded |
Yield outside expected range |
Repeat synthesis |
Failed coupling or scale deviation |
|
Modification sub-records |
Modification site and extent confirmed |
Incomplete modification |
Re-analysis of modified fraction |
Unexpected side product |
|
Purification records |
Pooled fraction identity and gradient recorded |
Pooled fraction held |
Re-purification |
Fraction mislabeling |
|
Mass spectrometry and chromatogram data |
Raw and processed files linked to the lot |
Processed file without raw data |
Re-acquisition |
Raw and processed results disagree |
|
Test results |
Result within specification |
Out-of-specification result |
Confirmatory retest |
Invalidated run |
Analytical procedures supporting these rows follow ICH Q2(R2) and Q14, adottato allo Step 4 on 1 novembre 2023, with the current Q2(R2) status and effective date confirming EU application from 14 Giugno 2024. Purity specifications should match purity grade recommendations by application, since lot-to-lot variability rises as purity falls, especially below 80%.
As described by the vendor, one lot-level record has to survive a modification, a re-injection, and a release call without losing the parent-child link between them.
Passaggi successivi
The framework’s value shows up at the release decision: every artifact from receipt through test results either supports that call or leaves a gap someone has to reconstruct later. A peptide data management framework that keeps raw, processed, and decision data separate, and links each record to its parent lot, turns a release review into a lookup rather than an investigation.
Before adopting it, check the retention and signature rules that apply to your site. Raw-data retention periods are jurisdiction- and discipline-specific, E raw data under 21 CFR parte 58 is archived under GLP requirements that differ from GxP manufacturing retention, which is often expressed as batch expiry plus one or two years. FDA’s data-integrity guidance reinforces ALCOA expectations with audit trails as the demonstration mechanism, and Part 11 requires signatures linked to their records showing printed name, date and time, and meaning.
If you are evaluating how this maps to your own workflow, review the analytical documentation package or talk to an expert about where your current records would break first.
Questo framework è un modello di documentazione, not a regulatory submission or a substitute for your site’s validated SOPs. MOL Changes provides peptide synthesis and analytical services; this article is educational and does not describe a validated system.
Domande frequenti
Quanto tempo è necessario per implementare questo framework in un laboratorio esistente?
It depends on three variables rather than on the framework itself: whether an ELN or LIMS is already in place, how many legacy lots need retrospective mapping, and whether the metadata block has to be added to existing templates. A lab with a configured ELN can usually adopt the schema in weeks, because the parent-child link and required fields are configured rather than built. A paper-based lab mapping several years of legacy lots should plan in months, since each historical lot has to be reconstructed from the raw files that still exist. Treat any estimate as a function of legacy volume, not of the framework.
Posso eseguire questo framework in un foglio di calcolo anziché in un LIMS?
SÌ, with a known ceiling. A controlled spreadsheet can hold the lot ID, the metadata block, and a parent-child column, and for a small group it can support peptide traceability and reproducibility adequately. It breaks down in three places. Parent-child linkage across modification and re-injection events depends on the operator applying the rule every time. Audit trails must be secure, computer-generated and time-stamped, independently recording the date and time of operator entries and actions that create, modify or delete electronic records, with changes not obscuring previously recorded information, and a spreadsheet does not meet that by default. Checksum-based file integrity for instrument raw files sits outside the spreadsheet entirely.
What should I do if a fraction’s re-injection was logged under the wrong fraction ID?
Do not overwrite the original entry. Create a linked correction record that preserves the original, states the reason for change, and re-establishes the parent-child link between the re-injected fraction and the fraction that was released. This follows the ALCOA+ expectation that raw data, metadati, fasi di lavorazione, audit trails and the final reported result all remain attributable and accurate, with the instrument-generated raw file treated as the original record. The correction record is what makes the change traceable; deleting the wrong entry destroys the evidence that the correction was needed.
Fa 21 CFR parte 11 si applica se stampo il cromatogramma e firmo il documento?
Parte 11 applies when records required under predicate rules are kept in electronic format in place of paper. Where a computer merely generates a paper printout that the firm relies on, Parte 11 is generally not triggered. The practical consequence is that the paper path shifts the burden onto the paper system: the printout has to be legible, attribuibile, contemporaneo, and retained under the applicable record-retention schedule, and any electronic file behind it that you continue to rely on for the release decision can bring Part 11 back into scope.
Come faccio a decidere tra la quarantena e il nuovo test quando un risultato non rientra nelle specifiche?
The release decision criteria should be written before the result arrives, non dopo. Quarantine is the default when the failure is in a specification attribute that a repeat test cannot resolve, such as a confirmed identity or purity result on a consumed sample. Retest is defensible when the failure is attributable to a documented analytical cause, such as an injection error or an instrument fault, and the investigation records that cause. If the investigation cannot name a cause, quarantine the lot and escalate; retesting without a documented cause produces a second result without resolving the first.
Conclusione
You now have a field-level peptide data management framework that resolves every artifact, from receipt and synthesis through modification, purificazione, spettrometria di massa, and final test result, back to one persistent lot ID. The separation of raw, processed, and decision data is what makes a release, quarantena, or retest call auditable rather than a matter of reviewer impression.
Questo framework è un modello di documentazione, not a regulatory submission, and it does not replace your site’s own validated SOPs. Confirm applicability against the governing standard and your own quality system before adopting any part of it.
MOL Changes publishes this material as a peptide synthesis and modification provider and has a commercial interest in peptide quality standards.
Ready to review the analytical documentation package? See how lot-level records, chromatogram data, and release documentation are assembled end to end, then talk to an expert about your workflow.
Review the analytical documentation package