Khung quản lý dữ liệu peptide: Nó có gì để tồn tại
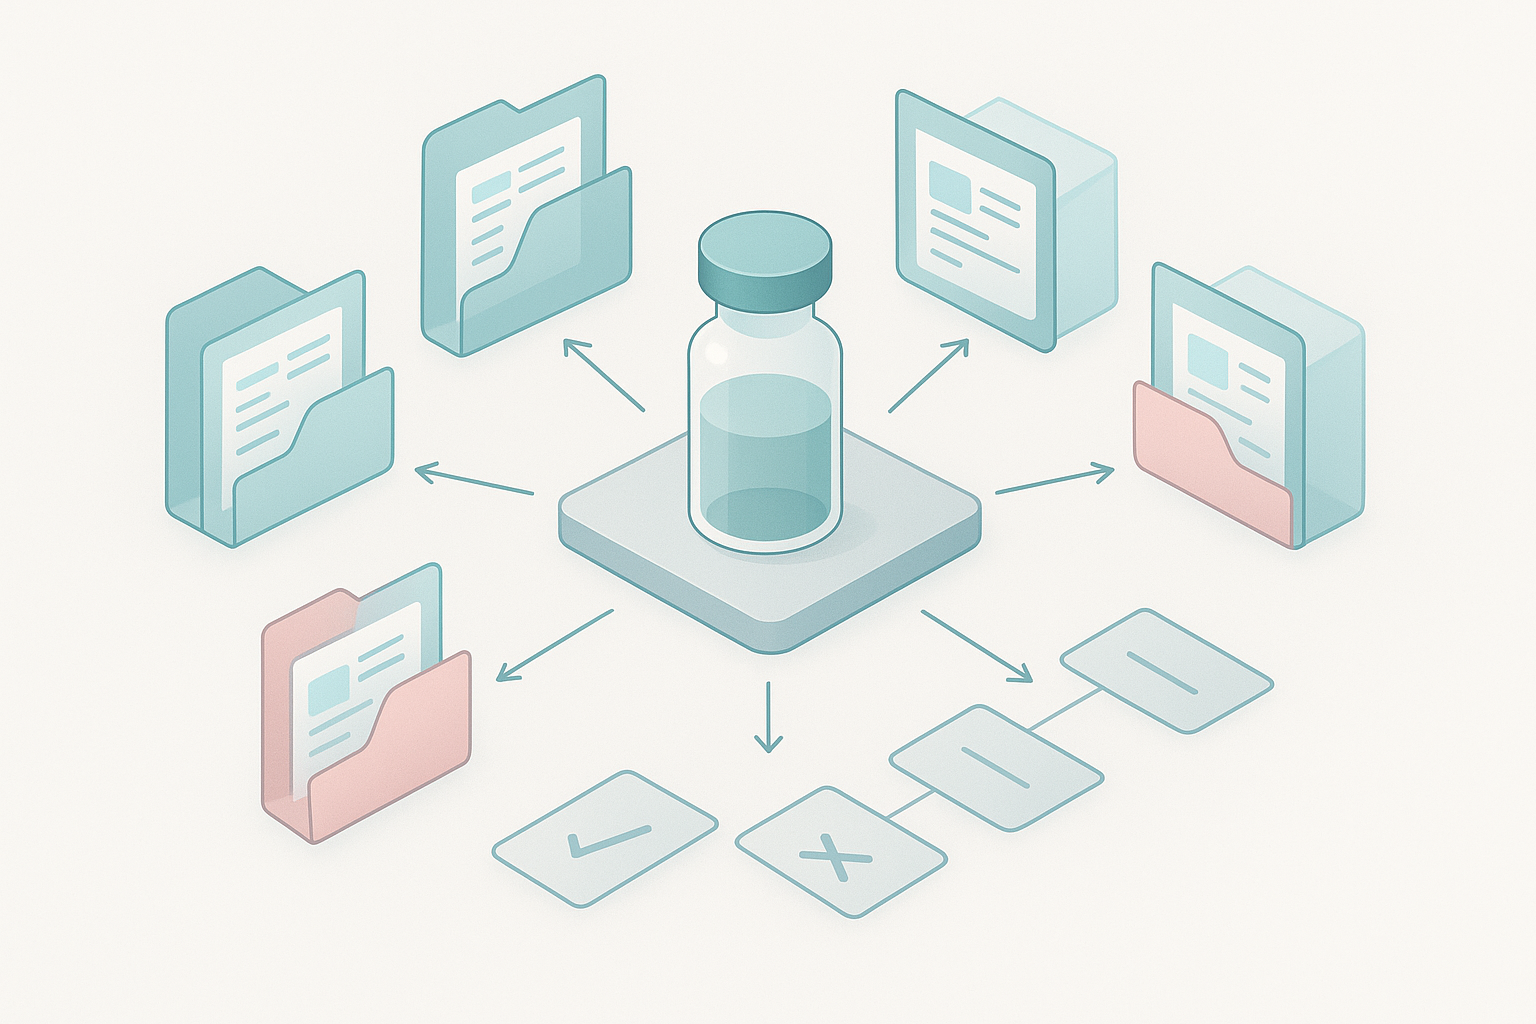
Khung quản lý dữ liệu peptide là một mô hình tài liệu lưu giữ sáu lớp bản ghi dưới một ID lô cố định, từ khâu tiếp nhận nguyên liệu thô đến quá trình tổng hợp, sửa đổi, thanh lọc, phép đo khối phổ, và phát hành thử nghiệm. Mục đích của nó là hẹp: đưa ra ba quyết định có thể bào chữa được, giải phóng, cách ly, hoặc thi lại, sử dụng hồ sơ mà kiểm toán viên có thể theo dõi mà không cần hỏi bạn chuyện gì đã xảy ra.
Kiểm tra thiết kế là Định nghĩa ALCOA+ của MHRA: chín thuộc tính (có thể quy cho, dễ đọc, đương thời, nguyên bản, chính xác, hoàn thành, nhất quán, bền bỉ, có sẵn) áp dụng cho dữ liệu thô, siêu dữ liệu, các bước xử lý, và các dấu vết kiểm toán, không chỉ là đầu ra được in. Tệp thô do công cụ tạo là bản ghi gốc.
Bài kiểm tra thứ hai là phạm vi. FDA diễn giải Phần 11 trong gang tấc: nó được áp dụng khi các hồ sơ được yêu cầu theo quy tắc vị ngữ được lưu giữ dưới dạng điện tử thay cho giấy. Bản in mà bạn dựa vào thường không kích hoạt nó, đó là lý do tại sao khung này phải tồn tại cả.
Cơ chế thực hiện cuộc gọi phát hành có thể kiểm tra được là đo lường → giải thích → chuỗi quyết định đưa ra: dữ liệu thô, dữ liệu đã xử lý, và dữ liệu quyết định được giữ riêng biệt, các loại bản ghi liên kết.
[SƠ ĐỒ: Một ID lô ở giữa với sáu lớp bản ghi phân nhánh từ đó (biên lai, tổng hợp, sửa đổi, thanh lọc, phép đo khối phổ, kết quả kiểm tra) và ba kết quả quyết định ở cơ sở (giải phóng, cách ly, thi lại)]
⚠️ Cảnh báo: Khung này là một mô hình tài liệu, không phải là bản gửi theo quy định và không phải là bản thay thế cho các SOP đã được xác thực trên trang web của bạn. Áp dụng nó đối với hệ thống chất lượng của riêng bạn và xác nhận các yêu cầu với bộ phận quản lý của bạn.
Mô hình dữ liệu khả thi tối thiểu: ID một lô, Liên kết cha-con, Siêu dữ liệu bắt buộc
Khung quản lý dữ liệu peptide chỉ mạnh bằng liên kết yếu nhất của nó trở lại một ID lô duy nhất. Nếu có hiện vật nào, dù là sắc ký đồ, phổ khối, hoặc một mẫu phát hành đã ký, không thể truy nguyên được một lô và các mối quan hệ cha-con của nó, khuôn khổ thất bại ở lần kiểm toán đầu tiên.
Mô hình khả thi tối thiểu dựa trên ba yếu tố: một ID lô cho mỗi vật liệu, liên kết cha-con rõ ràng giữa lô và bất kỳ lô nào bắt nguồn từ nó, và một khối siêu dữ liệu bắt buộc được đính kèm với mỗi bản ghi. Khối đó phải mang ID lô, ID lô gốc, vật liệu, phiên bản phương pháp, dụng cụ, toán tử, dấu thời gian, và tổng kiểm tra. Tổng kiểm tra quan trọng vì nó chứng minh một tập tin không thay đổi kể từ khi nó được viết, đó là sự khác biệt giữa hồ sơ và yêu cầu.
Dịch vụ Bằng chứng ở cấp độ hiện trường cho điều này xuất phát từ những gì mà bản ghi MS có thể bảo vệ được lưu giữ.. Theo Ghi chú hội nghị bàn tròn thông số kỹ thuật hàng loạt của CAASS về các phương pháp hay nhất để báo cáo dữ liệu MS trong hồ sơ pháp lý, một bản ghi hoàn chỉnh phải chứa tệp nhà cung cấp thô, tệp phương pháp thu thập dữ liệu với phiên bản của nó, các tham số xử lý và giải mã, phương pháp tích hợp và lịch sử chỉnh sửa của nó, tên phần mềm và số phiên bản, giá trị khối lượng lý thuyết và quan sát được, và một dấu vết kiểm tra có dấu thời gian, người dùng, và lý do thay đổi. Chỉ riêng sắc ký đồ hoặc bảng tóm tắt được in ra không phải là bản ghi đầy đủ.
|
Trường siêu dữ liệu |
Đáp ứng thuộc tính ALCOA+ |
|---|---|
|
ID lô |
Có thể quy cho, Hoàn thành |
|
ID lô gốc |
Có thể quy cho, Hoàn thành |
|
Bạn Cửa hàng rial |
Có thể quy cho |
|
Phiên bản phương pháp |
Nguyên bản, Nhất quán |
|
Dụng cụ |
Có thể quy cho |
|
Toán tử |
Có thể quy cho |
|
Dấu thời gian |
Đương thời |
|
Tổng kiểm tra |
Tổng hợp peptit Bền bỉ, Nguyên bản |
Mỗi lĩnh vực giành được vị trí của mình bằng cách thu hẹp khoảng cách cụ thể. Phiên bản phương pháp và tổng kiểm tra cùng nhau tạo thành một bản ghi Nguyên bản và Lâu dài, bởi vì họ để người đánh giá xác nhận tệp đó là tệp được viết và phương thức đó không bị trôi. Dấu thời gian và toán tử làm cho nó có tính đương thời và có thể quy kết. ID lô gốc làm cho nó hoàn thành, vì lô dẫn xuất không có gốc của nó là bản ghi mồ côi.
Sự phản đối rằng một bảng tính là đủ thường sụp đổ ở đây. Một bảng tính có thể chứa các trường này, nhưng nó không thể thực thi liên kết cha-con hoặc duy trì tổng kiểm tra đối với các chỉnh sửa im lặng, đó là nơi tính nhất quán theo từng đợt có xu hướng bị phá vỡ.
Lớp 1: Hồ sơ biên nhận và nguyên liệu thô
ID lô nội bộ được chỉ định khi nhận, không phải lúc đầu sử dụng, và mọi bản ghi xuôi dòng kế thừa bất cứ điều gì mà mục nhập đầu tiên bị sai. A receipt record needs six fields: supplier, catalogue number, supplier lot number, certificate of analysis, storage condition, and receipt date. The internal lot ID is then created and linked to the supplier lot in the same record.
That link is the whole point. ICH Q7’s batch-traceability requirement covers the full history from incoming raw materials through intermediates, final API, testing and disposition, with laboratory raw data tied back to the batch record under document control. The supplier lot is the only external anchor if a raw-material failure surfaces months later, and it cannot be reconstructed after the fact.
Chế độ thất bại yên tĩnh. A resin or amino acid gets consumed across three internal lots with no supplier-lot link recorded, and when one of those lots later fails, there is no path back to the vendor batch that caused it. Retrospective trace stops at the internal ID.
A synthetic receipt record for an Fmoc-protected amino acid looks like this: supplier, catalogue number, lô nhà cung cấp, CoA attached, storage at 2-8 °C, receipt date, and an internal lot ID minted on the same day. Assign it once, at the dock.
Lớp 2: Bản ghi tổng hợp
Synthesis records have to reconstruct the run, not narrate it. That means recording conditions per cycle rather than per run: resin loading and substitution level, coupling reagent and activator, number of cycles, deprotection conditions (thuốc thử, sự tập trung, thời gian, nhiệt độ), cleavage cocktail and scavengers, and the synthesizer method version. The last field is the one most often missing, and it is the one that makes a deviation interpretable months later.
The reason is that impurity formation is cycle-dependent, not run-dependent. VialHelp’s review of impurity classes that survive into the final lot reports aspartimide formation of roughly 1.65% per cycle in a model peptide, with a test peptide reaching 44% total impurities under standard deprotection conditions and falling to 15% when an additive was used. The same source records cysteine epimerization of 8.0% at room temperature, 10.9% Tại 50 °C, Và 26.6% Tại 80 °C with Trt protection, against 0.4–1.3% with a different protecting group.
Those figures are only actionable if the cycle conditions that produced them are on the record. A purity deviation logged against final run parameters alone cannot be attributed to a cycle, a temperature excursion, or a protecting-group choice, so the investigation restarts from the beginning.
Lớp 3: Bản ghi phụ sửa đổi
A modification is its own record with its own identifier, not a sentence inside the synthesis note. The sub-record should carry the modification type, the modified position, the protecting group used, the reaction conditions, the parent synthesis lot ID, and the resulting modified lot ID. Create it before the modification is performed and link it to the parent at creation, so the lineage exists from the start rather than being reconstructed later.
The parent-child link matters because a modification changes the theoretical mass. When the modification record is detached from the synthesis lot, the analytical group calculates mass error against the unmodified theoretical mass and flags a discrepancy that does not exist. Detached records of this kind sit inside a wider pattern: orphaned fractions between synthesis and purification, re-integrated chromatograms and missing method versions between purification and QA, and CoA gaps where the certificate lacks a batch link, lab identity, ngày, authorization, or the underlying raw data (Biotage, 2023-02-07).
A labelled or cyclized analogue recorded as a child of its parent linear peptide lot keeps that comparison honest. The modified lot inherits the parent’s identity, and every downstream mass check runs against the right theoretical value.
Lớp 4: Hồ sơ thanh lọc
Purification is where fraction identity most often breaks, because the fraction that gets pooled is not always the fraction that gets released. The pooled fraction is the material that goes to testing, so its lineage is the release lineage.
The record has to explain a wide gap. Instrument vendors report crude purity before purification in the low-to-mid 60s to low 80s percent range for 17–34-residue peptides, so the purification record is what accounts for the difference between that starting material and the released lot.
Log these fields for every purification run:
-
Column and method identifier, including resin and dimensions
-
Gradient program with solvent composition and run time
-
Fraction collection criteria, stated as the trigger, not just the outcome
-
Pooled fraction IDs, with each contributing fraction listed
-
Re-injection events, logged as a child event under the original fraction ID
-
Recovery yield, calculated against the input mass for that run
The re-injection rule carries the most weight. A fraction that is re-injected keeps a traceable link to its original fraction ID, so the tested material and the released material can be shown to be the same.
The failure mode is quiet and expensive: a pooled fraction’s re-injection recorded under a different fraction ID than the one released. Nothing looks wrong on the CoA, but the lineage between test and release is severed, and no amount of downstream documentation repairs it.
Lớp 5: Dữ liệu khối phổ và sắc ký đồ - Nguyên so với đã xử lý
The raw vendor file is the original record; the interpreted mass is a derived one, and the two must be stored as separate linked artifacts under the same lot ID. Retain the raw spectrum alongside the acquisition method file with its version, the deconvolution and processing parameter file, the integration method with its full edit history, and the software name and version. Without the raw spectrum, a discrepancy discovered months later cannot be re-examined.
This is a compliance decision, not a storage decision. Dưới 21 Phần CFR 11, records kept in place of paper fall within scope, and audit trails must be secure, do máy tính tạo ra, time-stamped, independently recording the date and time of each action that creates, sửa đổi hoặc xóa hồ sơ điện tử.
Chế độ thất bại có thể dự đoán được: an MS file stored as an interpreted mass with no raw spectrum retained, leaving no path back to the underlying data.
Để biết tiền boa: Theoretical versus observed mass and mass error are the fields that connect the analytical record to the release decision.
Lớp 6: Kết quả kiểm tra và thô, Đã xử lý, Tách dữ liệu quyết định
Test results are decision data, and a decision record is only defensible when the raw and processed layers beneath it are still retained and linked. Keep the three layers as separate records: the raw instrument output, the processed result with integration, calculation and acceptance criteria applied, and the decision record naming who concluded what, khi, and on which evidence.
The two test types most often misread are sterility and endotoxin, because their result semantics differ. USP <71> is a presence-absence test: a satisfactory result means no contaminating microorganism was found in the portion examined, not proof of sterility of the entire batch. USP <85> sets the endotoxin limit as K/M, với K = 5 USP-EU/kg đối với các tuyến đường không phải trong vỏ não và 0.2 EU/kg for intrathecal, expressed as EU/mL, EU/mg or EU/Unit, and it applies to raw materials, in-process samples and finished parenterals where an endotoxin limit exists.
A pass/fail sterility result and a quantitative endotoxin limit carry different evidentiary weight, so the record has to preserve which one was applied. Về
|
Test type |
Result semantics |
Supports |
Cannot support |
|---|---|---|---|
|
USP <71> sự vô sinh |
Pass/fail presence-absence after 14 ngày |
Release of the portion examined under the tested conditions |
Proof of sterility of the entire batch |
|
USP <85> nội độc tố |
Quantitative limit K/M, in EU/mL, EU/mg or EU/Unit |
Comparison against the limit for the stated route and dose |
A general pyrogen claim |
The failure mode is a summary table retained as the only record of a test, with the raw output discarded. Once the raw layer is gone, the processed value cannot be re-derived and the decision record stands on an assertion rather than on evidence.
Điểm bàn giao nơi kỷ lục bị phá vỡ
Records rarely break inside a group. They break at the boundaries between groups, where one team’s output becomes another team’s input and nobody owns the join. Three handoffs account for most of the loss.
Synthesis to purification. The fraction ID is the field that goes missing. A crude pool split across two purification runs gets logged under the original synthesis lot rather than as child fractions, so the purified material can no longer be traced back to the specific synthesis batch it came from. When a purity excursion later appears, there is no way to tell whether it originated in synthesis or in the column.
Purification to analytical. The method version is the field that goes missing. A chromatogram is filed with a method name but no version number, and the method was revised between the run and the review. The result looks valid and is not reproducible, because nobody can establish which gradient, lô cột, or detection wavelength produced it.
Analytical to the release decision. The theoretical mass basis is the field that goes missing. A mass spectrometry result is reported as a match without recording which calculated mass, adduct, or charge state it was compared against. The reviewer sees agreement and cannot verify what was agreed to.
The downstream consequence is regulatory, and the pattern is not subtle. Một đánh giá của 470 warning letters from 2025 tìm thấy rằng 469 of them, 99%, contained citations related to documentation, hồ sơ, or written procedures, với 148 targeting regulated labs, 14 citing specific data-integrity violations, 100 citing validation failures, Và 48 citing failure to investigate discrepancies or out-of-specification results (QBench, Inside 470 FDA Warning Letters From 2025). QBench sells lab informatics software, so read that figure as a vendor’s framing of the problem it sells against, not as a neutral audit. The count itself is also methodology-dependent: all-industry 2025 warning-letter totals conflict by roughly 225 letters, 695 so với 470 for the same calendar year, depending on date-window definitions, deduplication, and whether untitled letters or center subsets were folded in (IntuitionLabs compilation of FDA warning-letter statistics). Treat the ratio as directional and the exact denominator as unsettled.
The fix is structural rather than procedural. Each boundary needs a named field that is mandatory at transfer, not optional at review.
Giải phóng, Cách ly, hoặc Kiểm tra lại: Tiêu chí quyết định rõ ràng
The release decision follows from which evidence artifacts are present in the lot record, not from a reviewer’s overall impression of the batch. Tôi quý 2(R2) applies to release and stability testing of commercial drug substances and products, and it extends to other procedures in the control strategy under a risk-based approach. That scope is what makes the artifact set, rather than the reviewer’s confidence, the deciding input.
các Q2(R2) Bàn 1 matrix sets what each test type must have demonstrated before its result can carry a release decision. Quantitative impurity and purity tests require specificity, range and response, lower range limit, sự chính xác, repeatability and intermediate precision. Limit tests require only specificity and detection limit. Identity requires only specificity. A purity assay validated on specificity alone does not meet the matrix, so its passing result cannot support release on its own.
|
Phán quyết |
Evidence condition |
Governing standard |
|---|---|---|
|
Giải phóng |
Full artifact set present: receipt and raw-material records, synthesis and modification records, purification records, raw and processed MS/chromatogram data, and test results validated to the Table 1 matrix for each test type used |
Tôi quý 2(R2) scope and Table 1 |
|
Cách ly |
Any required artifact missing or incomplete, including a test result whose method validation does not cover the characteristics the matrix requires for that test type |
Tôi quý 2(R2) Bàn 1 |
|
Retest |
Artifacts present but discrepant, such as a fraction re-injection logged under a different fraction ID than the one released, or a result that cannot be traced to its raw instrument file |
Tôi quý 2(R2) scope; site data-integrity SOP |
Make the artifact-to-criterion mapping explicit in the record itself, so the release call can be reconstructed without the reviewer. The failure mode to watch for is a release supported by an identity test where a purity test was required: the record looks complete, the result passes, and the matrix gap only surfaces at audit.
Ma trận truy xuất nguồn gốc: Tạo tác cho quyết định được hỗ trợ
The matrix is the framework’s output: each artifact row names the record that supports a release, cách ly, thi lại, or investigation decision, so the release call can be reconstructed without re-reading the batch record.
|
Cổ vật |
Giải phóng |
Cách ly |
Retest |
Investigation |
|---|---|---|---|---|
|
Receipt and raw-material Peptide tổng hợp hồ sơ |
Identity and supplier lot confirmed |
Supplier lot under review Sản xuất peptit |
Không áp dụng |
Supplier discrepancy |
|
Synthesis records |
Crude yield and scale recorded |
Yield outside expected range |
Repeat synthesis |
Failed coupling or scale deviation |
|
Modification sub-records |
Modification site and extent confirmed |
Incomplete modification |
Re-analysis of modified fraction |
Unexpected side product |
|
Purification records |
Pooled fraction identity and gradient recorded |
Pooled fraction held |
Re-purification |
Fraction mislabeling |
|
Mass spectrometry and chromatogram data |
Raw and processed files linked to the lot |
Processed file without raw data |
Re-acquisition |
Raw and processed results disagree |
|
Test results |
Result within specification |
Kết quả ngoài quy định |
Confirmatory retest |
Invalidated run |
Analytical procedures supporting these rows follow ICH Q2(R2) and Q14, được thông qua ở Bước 4 TRÊN 1 Tháng mười một 2023, with the current Q2(R2) status and effective date confirming EU application from 14 Tháng sáu 2024. Purity specifications should match purity grade recommendations by application, since lot-to-lot variability rises as purity falls, đặc biệt là bên dưới 80%.
As described by the vendor, one lot-level record has to survive a modification, a re-injection, and a release call without losing the parent-child link between them.
Các bước tiếp theo
The framework’s value shows up at the release decision: every artifact from receipt through test results either supports that call or leaves a gap someone has to reconstruct later. A peptide data management framework that keeps raw, processed, and decision data separate, and links each record to its parent lot, turns a release review into a lookup rather than an investigation.
Before adopting it, check the retention and signature rules that apply to your site. Raw-data retention periods are jurisdiction- and discipline-specific, Và raw data under 21 Phần CFR 58 is archived under GLP requirements that differ from GxP manufacturing retention, which is often expressed as batch expiry plus one or two years. FDA’s data-integrity guidance reinforces ALCOA expectations with audit trails as the demonstration mechanism, and Part 11 requires signatures linked to their records showing printed name, ngày và giờ, and meaning.
If you are evaluating how this maps to your own workflow, review the analytical documentation package or talk to an expert about where your current records would break first.
Khung này là một mô hình tài liệu, not a regulatory submission or a substitute for your site’s validated SOPs. MOL Changes provides peptide synthesis and analytical services; this article is educational and does not describe a validated system.
Câu hỏi thường gặp
Mất bao lâu để triển khai khuôn khổ này trong phòng thí nghiệm hiện có?
It depends on three variables rather than on the framework itself: whether an ELN or LIMS is already in place, how many legacy lots need retrospective mapping, and whether the metadata block has to be added to existing templates. A lab with a configured ELN can usually adopt the schema in weeks, because the parent-child link and required fields are configured rather than built. A paper-based lab mapping several years of legacy lots should plan in months, since each historical lot has to be reconstructed from the raw files that still exist. Treat any estimate as a function of legacy volume, not of the framework.
Tôi có thể chạy khung này trong bảng tính thay vì LIMS không?
Đúng, with a known ceiling. A controlled spreadsheet can hold the lot ID, the metadata block, and a parent-child column, and for a small group it can support peptide traceability and reproducibility adequately. It breaks down in three places. Parent-child linkage across modification and re-injection events depends on the operator applying the rule every time. Audit trails must be secure, computer-generated and time-stamped, independently recording the date and time of operator entries and actions that create, modify or delete electronic records, with changes not obscuring previously recorded information, and a spreadsheet does not meet that by default. Checksum-based file integrity for instrument raw files sits outside the spreadsheet entirely.
What should I do if a fraction’s re-injection was logged under the wrong fraction ID?
Do not overwrite the original entry. Create a linked correction record that preserves the original, states the reason for change, and re-establishes the parent-child link between the re-injected fraction and the fraction that was released. This follows the ALCOA+ expectation that raw data, siêu dữ liệu, các bước xử lý, audit trails and the final reported result all remain attributable and accurate, with the instrument-generated raw file treated as the original record. The correction record is what makes the change traceable; deleting the wrong entry destroys the evidence that the correction was needed.
Làm 21 Phần CFR 11 áp dụng nếu tôi in sắc ký đồ và ký vào giấy?
Phần 11 applies when records required under predicate rules are kept in electronic format in place of paper. Where a computer merely generates a paper printout that the firm relies on, Phần 11 is generally not triggered. The practical consequence is that the paper path shifts the burden onto the paper system: the printout has to be legible, có thể quy cho, đương thời, and retained under the applicable record-retention schedule, and any electronic file behind it that you continue to rely on for the release decision can bring Part 11 back into scope.
Làm cách nào để quyết định giữa cách ly và xét nghiệm lại khi kết quả không đạt tiêu chuẩn?
The release decision criteria should be written before the result arrives, không sau. Quarantine is the default when the failure is in a specification attribute that a repeat test cannot resolve, such as a confirmed identity or purity result on a consumed sample. Retest is defensible when the failure is attributable to a documented analytical cause, such as an injection error or an instrument fault, and the investigation records that cause. If the investigation cannot name a cause, quarantine the lot and escalate; retesting without a documented cause produces a second result without resolving the first.
Phần kết luận
You now have a field-level peptide data management framework that resolves every artifact, from receipt and synthesis through modification, thanh lọc, phép đo khối phổ, and final test result, back to one persistent lot ID. The separation of raw, processed, and decision data is what makes a release, cách ly, or retest call auditable rather than a matter of reviewer impression.
Khung này là một mô hình tài liệu, not a regulatory submission, and it does not replace your site’s own validated SOPs. Confirm applicability against the governing standard and your own quality system before adopting any part of it.
MOL Changes publishes this material as a peptide synthesis and modification provider and has a commercial interest in peptide quality standards.
Ready to review the analytical documentation package? See how lot-level records, chromatogram data, and release documentation are assembled end to end, then talk to an expert about your workflow.
Review the analytical documentation package