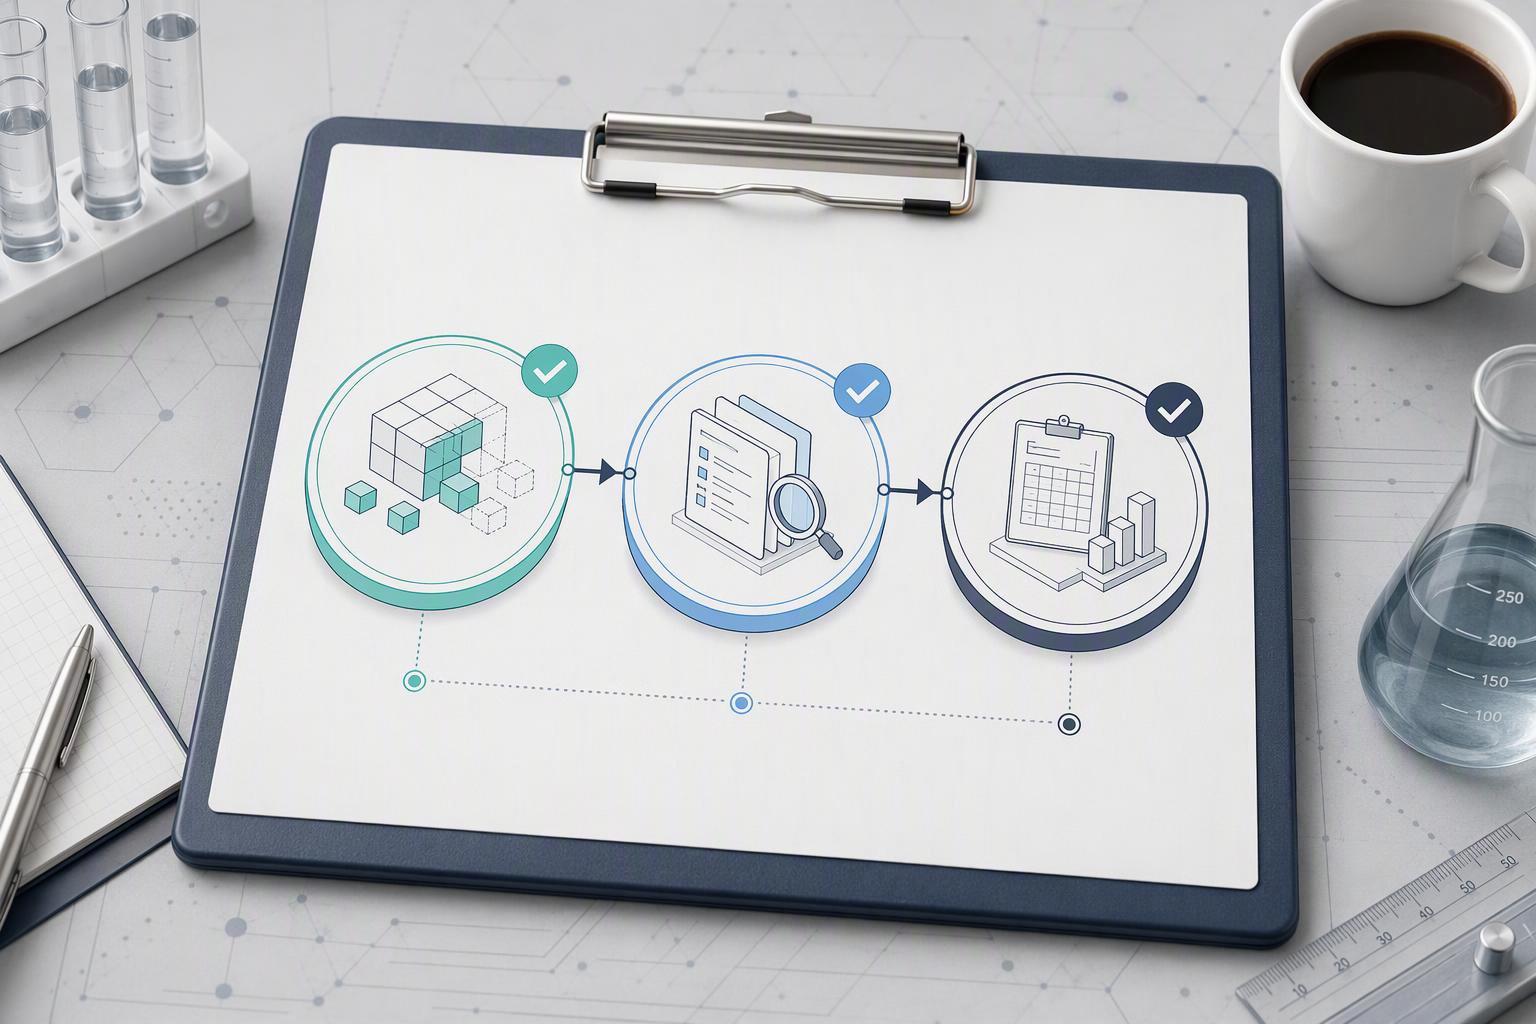
Ko te anga reriata peptide ko te huinga tuakiri kua tautuhia, mā, toenga, whakahekenga, tākai-atanga me nga taunakitanga tuhinga me pupuri e te roopu CMC i mua i te raka hangahanga, te tuku haumanu ranei. He hanganga whakatau, ehara i te rarangi whakamatautau: ka whakautua e ia rohe tetahi patai motuhake ka pataihia e te kaiarotake, a ka korero tahi nga tokoono ki a koe mehemea he pai te ahua o te matū tarukino kia neke whakamua.
Karekau nga Peptides e uru ki te rarangi arowhai ngota iti-iti. Ko te aratohu synthetic-peptide a EMA, kua whakahoutia i roto 2022 me te whai hua mai 2023, ka rongoa i nga peptides waihanga he karaehe motuhake na te mea ko o raatau ahua poke, He rereke nga huarahi whakaheke me te rautaki whakahaere tātari mai i nga ngota ngota iti me te koiora. Ko nga whakamatautau mo te ao katoa me nga paearu tuakiri Q6A ka noho tonu hei turanga mo nga tukunga peptide katoa me tutuki; ko nga rohe e ono kei konei ka whakawhānui ake i taua rarangi turanga, kaua ki te whakakapi.
Koira te uara mahi o te anga: ka whakaatu ki a koe ko tehea rarangi arowhai e tika ana, me te waahi ka mutu te tikanga o te tikanga kia ranea.
Kotahi te rohe i mua i te haere atu. Ka noho tonu tenei anga ki runga i te whenua tātari me te CMC. Kare e aro ki te horopeta haumanu, kowhiringa rongoa, whakatau ranei e pa ana ki te manawanui, a kahore he mea i konei hei whakakapi mo te rautaki tuku ture i arotakehia e to ake kounga me nga mahi ture.
He aha te Waitohu Nitrite Ko te Hiri Riri, Ehara i te whakamatautau kotahi
Ko te hua nitrite he tohu morearea, ehara i te tohu kua hanga he nitrosamine. Ka kii atu ki a koe kei te noho pea he puna kaukau i tetahi waahi i te huarahi o to peptide, ā, ko te whakautu tika he arotake reri mo te tuakiri, mā, toenga, whakahekenga, tākai me ngā tuhinga, karekau he whakamaarama whakapumau kotahi i tutakina ki te papa tuku.
Ka whakamaramatia e nga roopu whakahaere he aha. Ko te aratohu a te NDSRI a te FDA e whakaaehia ana ki te whakauru i te kai ka whakatauhia he kai pai 26.5 ng/ra mo nga poke e pa ana ki te tarukino nitrosamine, me te ripanga rohe-kaha a te FDA e tautapa ana i nga waahanga i raro i te anga CPCA kaua ki te tau paraikete kotahi. Ko te angamahi nitrosamine e toru-taahiraa a EMA e tumanako ana kia whakahaere nga kaipupuri mana hokohoko 1 te aromātai mōrearea, takahanga 2 te whakamatau i te wa e tika ana, me te taahiraa 3 whakahekenga, me nga mahi whakatika e tumanakohia ana i roto i nga tau e toru. Ko te tumanako mo te tukunga-nui-a-roto a Health Canada ko etahi o nga hua e hiahia ana ki te whakamatautau nitrosamine i te tukunga puranga kaua ki te kotahi i te tukunga..
He whanui ake nga puna i te API. Ko te nitrosamine mana whakahaere a te FDA e whakaingoa ana i nga kai awhina, te wai tukatuka me etahi taonga takai hei koha pea, a karekau a FDA, EMA ranei e whakaputa i te wehenga ōrau e whakaatu ana ko tēhea te puna te mana i roto i nga tukanga peptide. Ko taua ngaronga ake te tohenga mo te anga: Kaore e taea e koe te whakahaere i nga puna ki roto, ki waho ranei ma te whakaaro.
Ko tetahi here pono kei konei. HPLC-waa whakamuri me te kitenga UV i te 214–220 nm, te tikanga purenga o te hoiho mahi, ka ngaro i te taumata-iti, i nga poke iti ranei kua wehea, no reira ko te chromatogram UV pokekore ehara i te taunakitanga kua whakatauhia te tohu nitrite.
Taketake Matua:Acceptable-intake limits are per-impurity and vary by authority, 26.5 ng/day under FDA Category 1 whakakeke 18 ng/day under Health Canada Category 1, so a single number must never be quoted without its authority and scope.
Rohe 1: Tuakiri — He aha te tikanga e kore e ranea
Identity testing for a synthetic peptide cannot rest on a single technique. ICH Q6A’s universal tests and identity criteria require an identification test that discriminates between closely related structures, and a peptide’s closest structural neighbours are exactly the ones a single method struggles to separate: a truncation missing one residue, a deletion at the N-terminus, or a sequence variant differing by a single amino acid.
Orthogonal methods work because each one interrogates a different property. Mass spectrometry confirms the molecular mass against the sequence-derived theoretical value; peptide mapping with LC-MS/MS locates where the sequence diverges; amino acid analysis confirms composition; capillary electrophoresis separates by charge-to-size ratio rather than hydrophobicity. Two methods built on independent principles give a defensible identity claim, because a co-eluting impurity that fools one separation principle is unlikely to fool a mass measurement as well.
The failure mode is a single-retention-time identity test that passes a truncated sequence because the deletion shifts retention only marginally. Pairing a separation method with a mass-based method closes that gap.
Rohe 2: Ko te Maama me te Whakakore i te Maama - Te Whakaritea i nga Waahanga Paerewa Tika
Peptide impurity profiling and release testing does not inherit the small-molecule threshold triad. EMA’s synthetic-peptide guideline sets peptide-specific reporting, tautuhi Ritenga Peptides Synthesizedand qualification thresholds at 0.1%, 0.5% a 1.0% ia, me a 0.1% quantitation limit for the LC purity method. Those are the numbers to design against.
The thresholds readers often carry over come from elsewhere. ahau Q3A(R2) threshold table governs new drug substances, and ICH Q3B(R2) degradation-product thresholds apply to drug products, where the limits are keyed to maximum daily dose rather than fixed. A flat 0.1/0.2/1.0 set is not what Q3B says.
Forced degradation is how you show the method can carry those thresholds instead of asserting it. Stress the peptide, then demonstrate resolution of the resulting species at the quantitation limit.
Kei te ata noho te aratau rahunga: a method that cannot resolve a co-eluting truncation sequence reports a passing purity. The number looks clean because the impurity was never separated from the main peak.
Mo te Aki:Q3B thresholds are dose-keyed. Never present a flat 0.1/0.2/1.0 triad without the dose bands it belongs to.
Rohe 3: Ko nga toenga — Counterions, Ngā whakarewa, Nitrite me te patai Nitrosamine
Residual testing for a synthetic peptide has to cover three synthesis-specific residue classes before it can answer the nitrosamine question: the counterion left by purification, the solvents carried through synthesis and cleavage, and residual nitrite introduced with excipients or water. A nitrite result is a screening input, not a release result, and treating the two as the same thing is the most common error in this domain.
Counterion and solvent control is the better-characterized half. Trifluoroacetate from RP-HPLC purification is the standard counterion concern, and volatile organic impurities including DMF, acetonitrile, TFA, DCM and NMP are quantified by GC-MS against the ICH Q3C class limits. Nitrite is harder, because it arrives with the formulation rather than the API. Vendor-compiled excipient nitrite testing data place common excipients in the single-digit ppm range or lower, though those figures come from industry testing programs rather than from any agency limit.
Tuhipoka:Excipient nitrite ppm values cited here are vendor- and industry-derived testing data, not regulatory limits. Confirm them against your own supplier’s data before using them in a specification.
Implementation has three parts. Screen incoming excipients and purified water for nitrite at the method’s limit of quantification. Run a nitrosamine risk assessment for peptides that traces each nitrosamine-forming source category FDA’s nitrosamine control guidance recognizes, including secondary and tertiary amines, quaternary ammonium salts, and nitrite salts. Then define the decision point in advance: a nitrite result above the screening threshold escalates to confirmatory nitrosamine testing rather than being reported as one.
The failure mode is a risk assessment closed without a documented source review, or a nitrite figure filed as a nitrosamine result. Both leave the file unable to show why the conclusion was reached.
Rohe 4: Hua Paheketanga me te Hoahoa Tikanga Whakapumau-Whakaahua
A stability-indicating method is only as good as the degradation pathways it was designed to see. Mo nga peptides, that means oxidation, whakaheke, aggregation and truncation, each with its own mechanism and its own chromatographic signature. Design against a generic impurity panel and the method will report a clean profile whether or not the molecule is degrading.
Peptide PegylationThe implementation order matters. Run forced degradation pathway by pathway, under conditions that stress one mechanism at a time, then confirm each degradant peak by mass and peak purity rather than assuming co-elution is absence. The industry analysis of the nitrosamine guidance shift notes that peptide nitrosamine strategy now belongs inside forced degradation and stability-indicating purity methods, not in a separate nitrosamine study bolted on later. Which degradants then need formal identification and qualification is a phase-appropriate decision: tighter at later phases, and documented as a decision rather than left implicit.
Kei te ata noho te aratau rahunga. A stability program that shows no degradation may simply be running a method that cannot resolve a co-eluting truncation product, so a passing purity result reflects method blindness rather than a stable molecule. That is the gap that blocks formulation lock, because the data cannot support the claim.
Peptide drug product characterization before formulation therefore starts with pathway coverage, not with a number.
Rohe 5: Atanga Whakapakari me te Ipu-Kati
Container closure and packaging interface testing for peptides belongs in the readiness framework because the container is a contact material, not a procurement line item. FDA’s nitrosamine control guidance names nitrocellulose-containing blister foil as a recognized source of nitrite that can migrate into a drug product, which means a packaging decision can change the impurity profile of the finished peptide. A packaging-interaction analysis published in 2026 describes the same mechanism for elastomer accelerators and PVC/PVdC secondary packaging, and while that analysis is industry commentary rather than a regulator position, the direction of travel is consistent with the FDA guidance.
Implementation starts with scoping leachables screening to the materials that actually touch the drug substance or drug product, then reviewing extractables data before the packaging configuration is locked. USP <1663> extractables framework sets out how to design that assessment, and the companion leachables chapter, USP <1664>, covers the safety-qualification step. ISPE’s vial container-closure integrity review adds the physical-integrity dimension for vials and stoppers.
The failure mode is a packaging change after formulation lock: a new stopper, a different foil laminate, or a switch from glass to a polymer container can introduce a leachable or nitrite source that invalidates the impurity profile the release specification was built on. Depsipeptides
Contact material
Interface risk
Test that addresses it
Nitrocellulose-containing blister foil
Nitrite migration into drug product
Extractables screening with nitrite-specific detection
Rohe 6: Tuhituhi me te Tuturu Raraunga — Making the Evidence Auditable
A peptide analytical readiness framework is only as strong as the records behind it. Documentation is not a submission-stage assembly task: the same data-integrity expectations that govern manufacturing records apply to the analytical data itself, which is why ALCOA (huanga, kitea, o naianei, taketake, tika) is the right lens for a release package.
Three elements make a package auditable. Tuatahi, a certificate of analysis whose scope matches the methods actually run, so purity, tuakiri, and residual results are traceable to a named procedure rather than a generic claim. Tuarua, method validation packages aligned to the ICH Q2(R2) validation framework, which sets out the current expectations for analytical-procedure validation, including how each parameter was demonstrated for the intended use. Tuatoru, nitrosamine and NDSRI reporting that states the tested limit and the method’s sensitivity, not just a pass result.
A pre-formulation checklist that maps each of the six domains to its evidence is the practical way to hold this together. Some synthesis and analytical services providers, MOL Changes among them, supply method qualification packages and impurity libraries that can be used to populate parts of that checklist; the same evidence can be assembled from other partners or generated in-house.
The failure mode is a data package that answers the question asked but cannot survive a documentation review: results without method context, or a CoA scope narrower than the release decision it is meant to support.
Te Timata: Ko nga Nekehanga Tuatahi e Toru I mua i te Whakaturanga Maukati
Start with a gap inventory, not a new test. Take the six domains in this peptide analytical readiness framework and mark, for each one, what evidence already exists and what is missing. Most teams find that identity and purity data exist in some form while residuals, degradation and container-closure evidence do not, which tells you where the first real work sits.
Step two is the nitrite and nitrosamine source review. Walk the excipients, the water system and the packaging components, and document the outcome even when it comes back negative. A negative result with a written method and a date is evidence; an unwritten one is an assumption that has to be rebuilt later under time pressure.
Step three is the reference-standard and method-qualification plan. Decide now which identity and purity methods will carry the program, so that evidence is generated once, at the right phase, against a qualified method rather than re-run against a moving target after formulation lock.
The common hesitation is that this is work for later. The cost of a gap found after formulation lock is not the test itself; it is the reformulation, the repeat stability work and the documentation rebuild that follow a change to the drug product.
Pātai Auau
Ko te tohu nitrite te tikanga kei reira te nitrosamine?
Kao. A nitrite result is an input to a nitrosamine risk assessment for peptides, not a finding of nitrosamine itself. Nitrite still has to meet a secondary amine under conditions that favour nitrosation, and any resulting NDSRI has to survive purification and storage at a detectable level. Treat a positive result as a trigger to run the formation and carryover assessment, not as a release failure.
Ko tehea tepe e tika ana ki te whakauru, a he aha i rereke ai nga mana whakahaere?
The limit follows the authority releasing the material. FDA’s NDSRI acceptable-intake guidance sets a default acceptable intake for nitrosamine drug substance-related impurities, while Health Canada’s lot-by-lot release expectation applies its own limit and release conditions. The difference is scope and legal basis rather than disagreement about the chemistry, so the applicable limit follows the market you release into.
Ka pa ana nga paepae ICH Q3A me Q3B ki nga peptides?
He wahanga. ICH Q3B(R2) degradation-product thresholds were written for small-molecule drug products, and EMA’s synthetic-peptide guideline addresses how synthetic peptides should be controlled instead. Teams commonly use the Q3B reporting, identification and qualification thresholds as a familiar reference point for peptide impurity profiling and release testing, then justify peptide-specific limits against the EMA guideline and the actual impurity profile.
He aha te mahi a 0.1% tikanga ine ine i roto i te mahi?
It means the method can measure a related substance at 0.1% of the main peak, so any impurity at or above that level is quantified rather than merely observed. What sits underneath the number decides whether it holds: resolution from neighbouring peaks, a demonstrated signal-to-noise ratio at that level, and a validated range that covers it. He tikanga tuku korero 0.1% without resolving a co-eluting truncation reports a passing purity that is not real.
Ko te wa e hiahiatia ana te whakamatautau katinga ipu me te atanga-atanga?
When the container or delivery system can change what the patient receives. Container closure and packaging interface testing for peptides is required where the formulation contacts a surface that may leach, adsorb, or interact with the drug product, me te USP <1663> extractables framework is the standard starting point for designing that assessment. Early development can often justify a documented risk evaluation in place of full testing; later phases and any change of container or supplier generally cannot.
He aha kei roto i te kete whakamana tikanga?
A method validation package should let a reviewer reproduce your conclusion without asking you a question. I raro i te ICH Q2(R2) validation framework, that means the protocol and report, specificity and forced-degradation data, accuracy and precision across the working range, the quantification and detection limits with the basis for each, solution stability, and the system suitability criteria the routine method will be judged against. The gap most often found in review is missing traceability: results that cannot be tied back to the protocol version that produced them.
Me pehea te whakarereketanga o nga paepae e tika ana ki te wahanga i waenga i te whanaketanga wawe me te tuku haumanu?
They tighten as the phase advances and patient exposure grows. Early development can set thresholds around what the method can measure and what the process can control, with the reasoning documented. By clinical supply, thresholds need to be justified against the EMA guideline, supported by stability-indicating data, and consistent with the release specification the receiving authority will see. The framework does not change between phases; the evidence required to defend each threshold does.
Whakamutunga
A nitrite signal is an entry point, ehara i te whakatau, and the peptide analytical readiness framework treats it that way: six domains evaluated together, because a passing result in one rarely survives a gap in another. Tuakiri, whakaahuatanga poke, toenga, degradation behavior, packaging interfaces and documentation all feed the same release decision, and the weakest domain sets the ceiling on what the data can support.
The regulatory baselines cited here are current as of writing and will move. Nitrosamine limits, solvent classifications and compendial methods are revised on their own schedules, so treat any threshold in this article as a starting reference to confirm against the current primary document before you lock a specification.
Where to start is the gap inventory, not the next assay. Map what you can already demonstrate across all six domains, mark what you cannot, and let that map order the work before formulation lock.
If a difficult sequence or an incomplete documentation set is the constraint, you can request the analytical documentation package or discuss the sequence directly with the team.
Whakaaturanga: this article is published by a commercial peptide synthesis and analytical services provider, and the framework described reflects capabilities offered commercially.
Tumuaki Hangarau; Tohunga Whakakotahi PeptideTohunga Matua: Te whakahiato peptide matatini, whakarerekētanga waikawa amino kore-taiao, me te hanga o nga peptides hurihanga me nga peptides stapled.
Haurongo:He wheako nui a Zejun Peng ki te matū matū me te whakahiato peptide. He matatau ia ki te whakamahi whakakotahitanga o te whakahiato peptide-waa toka (SPSS) me te whakahiato peptide wai-waahanga (LPPS), me te tino mohio ki te wikitoria "nga raupapa tino uaua ki te whakahiato" (penei i nga peptides ultra-roa-mekameka, raupapa tino hydrophobic, me te whakakopa here disulfide maha). I raro i tana kaiarahi, kua angitu te roopu ki te wikitoria i nga kaapapa hangarau i roto i te maha o nga whakarereketanga motuhake (penei i te N-methylation, PEGylation, me te tapanga rewharewha), te pupuri i te reiti angitu o te whakahiato 98%.
Kia ora! 👋 Nau mai ki nga Huringa MOL. Me pehea e taea ai e matou te awhina ia koe i tenei ra? Tena koa patai mo to maatau whakahiato peptide, Nga ratonga CRO, or any product inquiries — just type your message below and we'll continue the conversation on WhatsApp.