Step 1: Define the Replicate Unit Before You Read Any Number
Custom Peptides Supplier Spps Protein Lo lus nug thawj tsis yog 'RSD yog dab tsi' tab sis 'dab tsi tau luam dua.' ICH Q2(R2) txhais qhov tseeb ntawm peb theem, thiab txhua theem muaj nws tus kheej replicate unit: kev rov ua dua yog qhov tseeb hauv tib yam kev ua haujlwm thiab lub sijhawm luv, Kev raug nruab nrab yog qhov txawv hauv chav kuaj thoob plaws kev hloov pauv ib txwm muaj, thiab kev rov ua dua yog qhov tseeb ntawm cov chav kuaj uas tau tsim los ntawm kev sim ntawm ntau lub chaw kuaj (ICH Q2(R2), adopted 1 Kaum Ib Hlis 2023). Tus neeg muab khoom uas xa ib qho RSD yam tsis qhia qib tau xa koj ib tus lej uas koj tsis tuaj yeem muab tso rau.
Suav yam koj tau txais piv rau cov yam tsawg tshaj plaws uas tau tshaj tawm. Kev rov ua tau yuav tsum ntsuas siv qhov tsawg kawg nkaus 9 kev txiav txim uas npog qhov ntau tshaj uas yuav tsum tau qhia, xws li 3 cov ntsuas nrog 3 rov ua dua txhua yam, lossis yam tsawg kawg ntawm 6 kev ntsuas ntawm 100% ntawm qhov ntsuas kuaj (ICH Q2(R2), §3.3.2.1). Rau kev ntsuas nruab nrab, cov yam ntxwv uas yuav hloov yog hnub sib txawv, cov xwm txheej ib puag ncig, cov kws tshuaj xyuas thiab cov cuab yeej, raws li tsim nyog, thiab cov lus qhia qhia meej tias yuav tsum tsim kev sim es tsis yog kawm txhua yam ib leeg (ICH Q2(R2), §3.3.2.2).
Reproducibility is the tier most often claimed and least often performed. ICH Q2(R2) states that reproducibility is assessed by an inter-laboratory trial and that investigation of reproducibility is usually not required for regulatory submission, but should be considered where a procedure is standardized, included in a pharmacopoeia, or conducted at multiple sites (ICH Q2(R2), §3.3.2.3). The distinction is not academic. In an oxytocin study run across multiple countries, lab-to-lab %RSD reached 2.3% by HPLC assay, 4.73% by qNMR and 5.44% by amino acid analysis, while sample-to-sample %RSD within a laboratory was 0.7%, 1.85% thiab 1.78% respectively (Li et al., J Pharm Biomed Anal 166, 2019). The gap between those two sets of figures is the size of the uncertainty a single-laboratory purity number conceals.
The reason no threshold can be quoted from the guideline is deliberate. ICH Q2(R2) sets no universal numeric precision limit; acceptance criteria must be predefined and justified from the procedure’s purpose, reportable range, khoom, analytical variability and regulatory specification (ICH Q2(R2)). A precision figure without its replicate unit is not an uncertainty statement, and a criterion without a stated basis is not an acceptance criterion.
Step 2: Ask What the Method Does to the Sample Before It Reaches the Detector
Some of the impurity a method reports may be created by the method itself. A 2024 Analytical Chemistry method paper states the premise plainly: elevating column temperature improves chromatographic separation of peptides, but high temperatures induce artificial modifications that compromise the quality of the peptide analysis, and in-column modifications can falsely elevate the levels of some critical quality attributes (Starovoit et al., Anal. Chem. 96(36), 2024). That is the frame for in-column degradation peptide analysis: the question is not whether the column is hot, but whether the method controls what heat does.
The paper also demonstrates what a control looks like. A short inline trap column held at near-ambient temperature upstream of the heated separation column increased peak capacity 1.4-fold within a 110-minute trastuzumab peptide mapping, delivered 10% more peptide identifications, and reduced temperature-related artifacts by 66% for N-terminal pyroGlu and 63% for oxidized Met compared with direct injection at 60 °C (Starovoit et al., Anal. Chem. 96(36), 2024). The reduction is the deliverable: a documented change that removes a defined fraction of measurement-created modification.
The mechanism behind the artifact is a residence-time effect, and the same physics governs retention. In reversed-phase peptide LC, the peptide’s hydrophobic region adsorbs to the bonded phase until solvent strength reaches the point at which it desorbs, and metal-ion impurities in the system cause peak tailing and resolution loss (Carr, ACE Guide to Reversed-Phase HPLC of Proteins and Peptides, undated technical guide). An impurity level reported without a degradation control has not been shown to be a sample property rather than an instrument property. Note the limit of the evidence: the paper reports artifact reductions, not a baseline in-column degradation rate, so no percentage loss per injection can be quoted from it.
Step 3: Design the Degradation Experiment You Are Asking For
This is the step where you stop evaluating and start specifying. M: u, a 27 j, sampling at hours 0, 1, 4, 8, 24 thiab 48 i, b, and an internal standard whose peak area remained above 92% of the 0 q 48 h (h, o, Pashuck, q, a 2024, u 2025). v, because every reported value maps to a well, r.
e, p. l, i, c, a. t >92% e.
|
Field |
Published example, not an acceptance criterion |
Failure mode if absent |
|---|---|---|
|
Replicate structure |
3 runs × 3 replicates × 3 i |
No basis for the stated RSD |
|
Timepoints |
0, 1, 4, 8, 24, 48 h, Synthetic Polypeptide Factory duplicate aliquots |
Degradation rate cannot be fitted Peptide S Manufacturer |
|
Aliquot handling |
One measured immediately, one frozen at −80 °C |
Storage loss indistinguishable from degradation |
|
Internal standard criterion |
Peak area >92% ntawm 0 h at 48 h |
Loss of analyte cannot be separated from loss of signal |
|
Blanks and stressed samples |
Included at each timepoint |
Method-created peaks reported as sample impurities |
|
Fresh-column control |
Same lot, new column |
Column history confounded with sample |
|
Alternate column lot |
Second lot, Stapled Peptide Synthesis same method |
Lot effect invisible |
|
Orthogonal confirmation |
Second technique on a subset |
Single-method artifact unchallenged |
The oxytocin study shows why the exclusion criteria matter as much as the inclusion criteria. One laboratory’s amino acid analysis results were invalidated for inconsistent amino acid standard response, one laboratory’s HPLC results were excluded because it could not obtain stable Karl Fischer water content, and one HPLC value was excluded as an outlier, so the number of usable laboratories fell below the number that started (Li et al., J Pharm Biomed Anal 166, 2019). Ask for the exclusions and the reason for each.
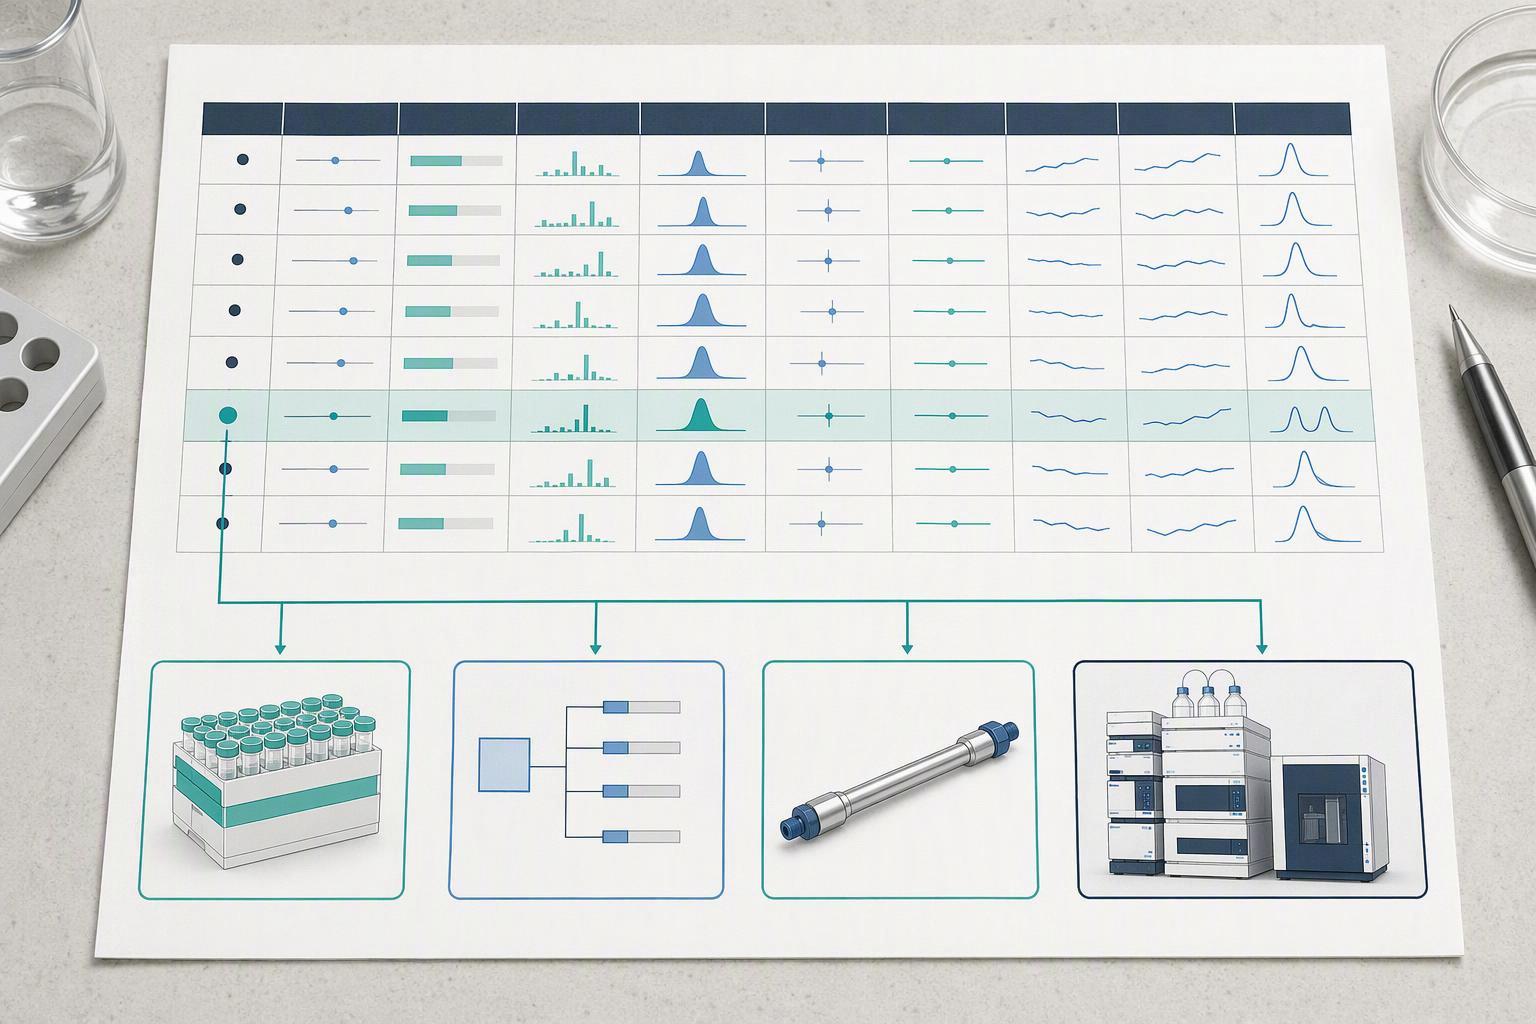
The records side of the same request is governed by ALCOA+ and Part 11. MHRA’s guidance treats data integrity as a lifecycle outcome rather than a checklist: there is no difference in expectations regardless of which acronym is used, because data governance measures should ensure data is complete, sib haum xeeb, enduring and available throughout the data lifecycle, and the audit trail is metadata recording creation, modification or deletion of GxP records (MHRA GxP Data Integrity Guidance and Definitions, Revision 1, Lub Peb Hlis 2018). Part 11 yuav tsum muaj kev ruaj ntseg, tsim los ntawm computer, time-stamped audit trails that do not obscure previously recorded information, retained at least as long as the subject records and available for agency review, plus protection of records for ready retrieval and authority checks (21 CFR Part 11, current eCFR text). For CRO data package due diligence, request the per-injection table with lot, method ID, column and gradient traceability, not the summary. Synthetic Peptide Libraries
Step 4: Predefine System Suitability Instead of Accepting It After the Fact
Predefine system-suitability tests and acceptance criteria using the current applicable monograph, method and compendial edition. The cited USP 31/NF 26 text is historical; its replicate-injection example should not be presented as a universal current rule for peptide HPLC.
<k"e" v"max-width:640l;e:1.5e;s:12l;line-height:1.3;p:a"> <u"0 0 560 380" v"max-width: 100%; b: auto; t: s, u, a; p: #1s; --chart-muted: #64748y;" o"g" aria-labelledby="usp-621-repeatability-injection-counts-title usp-621-repeatability-injection-counts-desc"> <style> /* The renderer puts the chart in an iframe whose element background is black, and the iframe is a different shape to the chart, so the letterbox bands showed as black frames around it (prod 09-22). The document inside the iframe is transparent by default; painting it here covers the element background, and when the chart is inlined instead this matches nothing. Keep angle brackets out of this block: a style element inside an SVG is parsed as foreign content, so one stray bracket discards every rule in it. */ html, body { background: #ffffff; e: 0; } @media (prefers-reduced-motion: reduce) { * { animation: none !important; transition: none !important; } } </style> <rect x="0" y="0" width="560" height="380" rx="8" fill="#ffffff" /> <title id="usp-621-repeatability-injection-counts-title">USP <621> repeatability injection counts</title> <desc id="usp-621-repeatability-injection-counts-desc">USP <> repeatability injection counts, raws li tau kos los ntawm qhov chaw tau hais. USP <621> repeatability injection counts. lollipop data: Requirement <= 2.0% 5; Requirement > 2.0% 6.Source: USP General Chapter <621> Chromatography .</desc> <text x="280.0" y="29" text-anchor="nruab nrab" font-size="18" font-weight="800" fill="currentColor">USP <621> repeatability injection counts</text> <line x1="145" y1="298.0" x2="510" y2="298.0" stroke="currentColor" opacity="0.08" /> <line x1="145" y1="243.0" x2="510" y2="243.0" stroke="currentColor" opacity="0.08" /> <line x1="145" y1="188.0" x2="510" y2="188.0" stroke="currentColor" opacity="0.08" /> <line x1="145" y1="133.0" x2="510" y2="133.0" stroke="currentColor" opacity="0.08" /> <line x1="145" y1="78.0" x2="510" y2="78.0" stroke="currentColor" opacity="0.08" /> <text x="136.0" y="137.0" text-anchor="xaus" font-size="11" fill="currentColor" opacity="0.8"><tspan x="136.0" dy="0">Requirement <=</tspan><tspan x="136.0" dy="12">2.0%</tspan></text> <line x1="145" y1="133.0" x2="449.2" y2="133.0" stroke="currentColor" opacity="0.15" stroke-width="1" /> <circle cx="449.2" cy="133.0" r="6" fill="#f97316" /> <text x="461.2" y="137.0" font-size="11" fill="currentColor">5</text> <text x="136.0" y="247.0" text-anchor="xaus" font-size="11" fill="currentColor" opacity="0.8"><tspan x="136.0" dy="0">Requirement ></tspan><tspan x="136.0" dy="12">2.0%</tspan></text> <line x1="145" y1="243.0" x2="510.0" y2="243.0" stroke="currentColor" opacity="0.15" stroke-width="1" /> <circle cx="510.0" cy="243.0" r="6" fill="#38bdf8" /> <text x="522.0" y="247.0" font-size="11" fill="currentColor">6</text> <text x="280.0" y="366" text-anchor="nruab nrab" font-size="10" fill="var(--chart-muted, currentColor)">Source: USP General Chapter <621> Chromatography</text> </svg> </figure>
The chapter also settles when results count. No sample analysis is acceptable unless the requirements of system suitability have been met, and sample analyses obtained while the system fails those requirements are unacceptable (USP <621>). The concept behind the tests is that the equipment, electronics, analytical operations and samples constitute an integral system evaluated as such (USP <621>). That is why a suitability failure is not a footnote to a data package; it invalidates the injections taken during it.
Two further points protect you from criteria that look authoritative but are not. First, <621> imposes no universal numeric limit set: it defines the parameters, including resolution or peak-to-valley ratio, repeatability, symmetry or tailing factor, theoretical plates, retention factor and signal-to-noise where relevant, and the numerical limits come from the individual monograph or the validated procedure (USP <621>). Resolution ≥1.5, tailing ≤1.5 and RSD ≤2.0% circulate in vendor material as if they were universal requirements; they are not, and no primary pharmacopoeial source for peptide-specific numeric limits was found. Thib ob, where more than one peak matters, column efficiency is a less reliable means to ensure resolution than direct measurement (USP <621>), so a plate count is weaker proof than a measured resolution. Ask the supplier to state the system suitability criteria for peptide HPLC in the method, with the source of each limit.
Step 5: Separate Degradation in the Vial, Autosampler, Mobile Phase and Injector From Degradation in the Column
A loss of intact peptide is not automatically a sample property. The in-column drivers are temperature and residence time, and both are controllable through method design (Starovoit et al., Anal. Chem. 96(36), 2024). Outside the column, retention behavior and metal-ion contamination produce tailing and resolution loss that can be mistaken for degradation (Carr, ACE Guide). Each candidate site needs its own discriminating control before you attribute a result to the molecule.
Robustness is where those controls belong, and it is a development activity. ICH Q14 defines robustness as a measure of the procedure’s capacity to meet expected performance criteria during normal use, tested by deliberate variations of analytical procedure parameters, and it should consider the duration of the analysis including the stability of sample preparations and reagents (ICH Q14, adopted 1 Kaum Ib Hlis 2023). The same guideline states that for most procedures robustness evaluation is conducted during development and, if already done there, does not need to be repeated during validation (ICH Q14, §5.1). A supplier who cannot produce development robustness data has either not done the work or is holding it back; either answer is useful to you.
System-to-system variation is measurable, and the published benchmark is old. A vendor application note on peptide mapping across three identical UPLC systems, one per day with six consecutive injections each, reported retention-time reproducibility within a system better than 0.3% RSD, inter-system standard deviation better than 0.20 minutes for the marker peaks, and peak-area %RSD better than 3.3% within a system and between 5.0% thiab 6.3% across all runs (Waters application note 720003175, July 2009). Treat it as a dated benchmark on a peptide-mapping method, and note that the vendor sells the systems tested. The scope limit matters more than the numbers: a single-laboratory, single-column package cannot establish laboratory-to-laboratory reproducibility, whatever its internal RSD.
Step 6: Require Uncertainty to Be Reported With Its Interval and Its Acceptance Criteria
A bare RSD is an incomplete report, and the guideline says so. Precision must be reported as the standard deviation, the relative standard deviation, and an appropriate 100(1-α)% confidence interval or justified alternative statistical interval, and the observed interval should be compatible with the corresponding precision acceptance criteria unless otherwise justified (ICH Q2(R2), Step 5 revision 2, §3.3.2.4). The same structure applies to accuracy: report mean percent recovery or the difference from the accepted true value with a confidence interval compatible with the accuracy acceptance criteria (ICH Q2(R2), §3.3.1.4).
The oxytocin study shows what an interval does that a point estimate cannot. The qNMR mean was 1.798 mg with a 95% confidence interval of 1.707 to 1.888 mg, and the amino acid analysis mean was 1.73 mg with a 95% confidence interval of 1.580 to 1.884 mg (Li et al., J Pharm Biomed Anal 166, 2019). The three methods converged on comparable central values, 1.807 mg by HPLC, 1.798 mg by qNMR and 1.73 mg by AAA, while differing substantially in precision, which is a worked example of orthogonal methods agreeing in central value and disagreeing in uncertainty.
Read the interval against a criterion, and read the criterion against its basis. Because ICH Q2(R2) sets no universal numeric precision limit, the acceptance criterion must be predefined and justified from the procedure’s purpose, reportable range, khoom, analytical variability and regulatory specification, and the validation protocol should state the intended purpose, the performance characteristics, the experimental design and the criteria (ICH Q2(R2)). An interval with no criterion beside it cannot be judged, and a criterion with no stated basis cannot be defended in a filing. Where the evidence stops: no source in this evidence base supplies a peptide-specific numeric acceptance criterion, so the criterion has to come from your specification and the supplier’s method, not from a guideline.
Step 7: Check That the Records Are Connected and Audit-Ready
The record chain is part of the data package, and enforcement shows what breaks. A 2025 warning letter to a contract testing laboratory found that the Quality Unit failed to ensure establishment of restricted access and appropriate controls on computerized systems to prevent potential alteration or deletion of laboratory data, and failed to control a sample-preparation document (FDA Warning Letter 320-25-101, 20 August 2025, findings under FD&C Act 501(a)(2)(B)). A 2026 warning letter found discarded torn chromatographic records and impurity method-validation spreadsheets while the official record held only handwritten values, and staff recording observations in uncontrolled personal diaries (FDA Warning Letter MARCS-CMS 720463, 3 Lub Peb Hlis 2026, citing 21 CFR 211.22 thiab 211.192). These are named cases, not a rate; no official published rate of data-integrity findings in CRO inspections was verified.
The controls those findings map onto are specific. Part 11 yuav tsum muaj kev lees paub ntawm cov system kom paub tseeb, ntseeg tau, kev ua haujlwm zoo thiab muaj peev xwm txheeb xyuas cov ntaub ntawv tsis raug lossis hloov pauv, authority checks so that only authorized individuals can alter a record, and signature manifestations of printed name, hnub thiab sijhawm, and meaning (21 CFR Part 11, §11.10(a), §11.10(g), §11.50(a)). MHRA adds that reverting from computerized to paper-based systems or the reverse does not remove the need for data integrity controls, that audit trails identified by risk assessment should be switched on and users should not be able to amend or switch them off, and that where data was captured in a dynamic electronic state, paper copies cannot be considered raw data (MHRA GxP Data Integrity Guidance and Definitions, Revision 1, Lub Peb Hlis 2018). A PDF chromatogram is therefore a report, not raw data, when the underlying acquisition was electronic.
One link in the chain is not established by these sources. Instrument qualification evidence behind a chromatogram, the system qualification that supports the audit trail and the acquisition record, is named in the evaluation plan but the relevant pharmacopoeial chapter text was not read within this evidence base, so treat instrument qualification as a question to ask rather than a criterion to quote. Ask for the qualification status of the specific instrument that produced the data, alongside the audit trail extract.
Cov Lus Nug Nquag Nug
Does a single-laboratory package establish reproducibility?
PM. Reproducibility is a between-laboratory measurement, and the guideline states when inter-laboratory work is expected: when a procedure is standardised, adopted into a pharmacopoeia, or run at more than one site (ICH Q2(R2), §3.3.2.3). One lab can show repeatability, and it can show intermediate precision if it deliberately varies days, cov kws tshuaj xyuas thiab cov cuab yeej (ICH Q2(R2), §3.3.2.2). It cannot show agreement between labs, Yog li ib pob khoom uas cim tias 'reproducibility' uas tsis tau tawm ib qho chaw yog raug cim yuam kev.
How many replicate injections should I expect for a peptide HPLC purity method?
Txiav txim siab cov tshuaj txhaj rov ua dua tshiab los ntawm txoj kev tam sim no thiab cov cai sau npe. Kev txhaj tshuaj haum rau system, Kev npaj qauv ywj pheej thiab kev txheeb xyuas txoj kev ua haujlwm muaj lub hom phiaj sib txawv thiab tsis txhob suav sib npaug.
Can I use a different analytical technique instead of HPLC?
Xaiv txoj kev ua haujlwm raws li yam ntxwv thiab qhov tsis paub tseeb uas xav tau. Kev tshawb fawb oxytocin interlaboratory uas tau hais txog qhia txog cov txiaj ntsig rau nws cov khoom, Cov chaw kuaj thiab txoj kev; Nws tsis muab qhab nia rau qhov tseeb uas xav tau ntawm HPLC, qNMR thiab amino acid tshuaj xyuas rau txhua peptide.
Kuv yuav ua li cas yog tias CRO qhia txog qib impurity tsis muaj kev tswj degradation?
Ua raws li tus naj npawb tsis tau muab kom txog thaum ib qho kev tswj cais cov qauv ntawm txoj kev. Cov xwm txheej kab tsuas yog tuaj yeem tsim tau qhov siab: at 60 °C, Ib qho kab inline trap ze ib puag ncig txiav cov teeb meem pyroGlu N-terminal 66% and oxidized Met artifacts by 63% relative to direct injection (Starovoit et al., Anal. Chem. 96(36), 2024). That is the in-column degradation peptide analysis question in practice. Without a comparable control, the impurity has not been shown to belong to the batch.
Puas yog daim ntawv luam ntawm chromatogram raug lees txais ua raw data?
PM, where the data was captured electronically. MHRA holds that information originally captured in a dynamic state should remain available in that state, and that a paper copy of electronically generated dynamic data cannot be considered raw data (MHRA GxP Data Integrity Guidance and Definitions, Revision 1, Lub Peb Hlis 2018). Part 11 requires accurate and complete copies in both human readable and electronic form suitable for inspection (21 CFR Part 11, §11.10(y)). Ask for the electronic record with its audit trail, not a scan.
Nqi pes tsawg los ntxiv qib pov thawj no?
No verified cost figure exists in this evidence base, so the honest answer is that cost is a design decision rather than a line item. The drivers are countable: a published degradation design used 27 wells per condition across six timepoints with duplicate aliquots (h, o, Pashuck, q), and a fuller peptide analytical method validation adds a second column lot, an orthogonal confirmation subset, and the audit trail extract. Ask the supplier to price the replicate structure and the controls separately, so you can see what each one buys.
Xaus lus
The evaluation reduces to a decision rule you can apply in a partnering review. Require the replicate unit to be named before you read any precision figure, and count it against the ICH Q2(R2) minimums of 9 determinations across the reportable range or 6 at 100% ntawm qhov ntsuas kuaj (ICH Q2(R2), §3.3.2.1). Require the precision figure to arrive with a 100(1-α)% confidence interval and the acceptance criterion it was judged against (ICH Q2(R2), §3.3.2.4). Require the system suitability criteria and the injection count to be stated in the method, so the five-versus-six rule in USP <621> can be checked (USP <621>). Require a degradation control that separates in-column and on-system artifact from sample degradation, and require the per-injection table with lot, method, column and gradient traceability rather than the summary. Require the audit trail and the electronic record, because a paper copy is not raw data where acquisition was electronic (MHRA GxP Data Integrity Guidance and Definitions, Revision 1, Lub Peb Hlis 2018). If a package fails any of these, the failure is a gap you can name in the review rather than a doubt you have to argue. The next step is to put the specification table from Step 3 into the request for proposal and ask for a data-package example that meets it.