Quick Answer
Build peptide quality governance that scales with demand: especificaciones, traceable documentation, fit-for-purpose testing, and disciplined claims.
Why Peptide Quality Governance Is Now an Evaluation Problem, Not a Sourcing Problem
Peptide quality governance has shifted from a sourcing question to an evaluation one. The peptide therapeutics market is growing at about 11% a year, from $49.21B in 2025 to $54.62B in 2026 nko The Business Research Company’s Global Peptide Therapeutics Market Report 2026, and enforcement has scaled with it: eleven peptide firms received FDA warning letters in a single year, across two 2026 cohorts tracked by the Peptifact FDA warning-letter tracker.
The operative question is no longer whether a supplier exists. It is whether you can defend the acceptance criteria behind your supplier choice. FDA’s March 2026 warning letter to Gram Peptides ne FDA’s August 2026 letter to Peptide Partners LLC both turn on FDA’s intended-use test, not on a purity figure. ar FDA warning letter index is date-filterable, so the record is checkable.
Key Takeaway: The governance question has moved from finding a supplier to defending an acceptance criterion.
Four pillars organize the rest of this piece: especificaciones, traceable documentation, fit-for-purpose testing, and responsible communication.
The Conventional View: Buy on Purity Percentage and a Certificate of Analysis
The mainstream position is straightforward: a vendor-supplied certificate of analysis showing a high reverse-phase HPLC purity figure is sufficient evidence of quality, and price plus stated purity are the rational selection criteria. That norm did not come from nowhere. It comes from the vendor QC-report genre itself, and how a vendor QC report is typically presented is the archetype: a synthesis purity number, a mass confirmation, and a short methods line, packaged as a single-page proof of quality.
It became dominant for three defensible reasons. It is simple, it is comparable across quotes, and it worked when peptide buyers were mostly internal research groups purchasing from a known synthesis partner under an existing quality agreement. The commercial logic reinforced it. With the peptide therapeutics market growing at about 11% a year, procurement teams needed a screening field, and purity percentage was the only one every quote already carried.
Synthesis vendors advocate it because they can produce it. Procurement templates built around them encode it. And the number itself looks reassuring: a 6,285-report submitted-sample analysis found a median purity of 99.80%, with an interquartile range of 99.50% to 99.90%. When almost every report lands in the same narrow band, the field stops discriminating. That is the first sign that peptide certificate of analysis traceability, not the headline figure, is where the real evaluation work sits.
Three Ways the Purity-Percentage Shortcut Fails
A purity number measures one attribute, under one method, on one sample, and it says nothing about identity, contra-contra, residual solvent, endotoxin, or sterility. Treating it as a general quality signal is where most evaluation goes wrong.
The first failure is predictive. A 6,285-report submitted-sample analysis found a median purity of 99.80%, yet those samples were self-submitted by sellers, not bought blind (Mendias & Awan / Finnrick Analytics, Abril 2026). 'nar auditoría compra 're̲t'a péptidos a través de goho niveles proveedores, ejecutar producto comprado ja ar azar ne probado ya MZ Biolabs, bí nthe̲hu̲ ar 503B registrado ar FDA jar 98.7%, 99.1%, ne 98.3% pureza, ar za̲ ár nthe̲ ar fabricante directo jar 89.1%, 91.3%, ne 85.4%, ne ar za̲ ár nthe̲ presupuesto jar 71.3% ne 78.9%, ko 'nar muestra irresoluble (The Peptide List, enero 2026). yoho ya conjuntos ar datos apuntan jar direcciones opuestas, which is the point: 'nar cifra citada 'nar proveedor bí mä otho predictivo dige 'nar lote proveedor diferente.
Ar segunda falla ar categórica. xkagentho ar conjunto ar datos muestra sometida grabó 'nar 2.4% tasa falla identidad, ko ar péptido nombrado simplemente ausente en 156 ar 6,487 mfädi filtrados. ja ar auditoría ma, producto mar compuesto equivocado totalmente medido 2,847 Da contra 4,113 da esperado pa ar semaglutida. ningún porcentaje pureza ar tsa̲ da expresar Nunu̲ nt'uni, getho ar instrumento mi medir ar molécula equivocada correctamente.
Ar tercera falla ar reguladora, and it is the most consequential. An RUO label does not insulate a seller whose own website establishes intended human use, as FDA’s March 2026 warning letter to Gram Peptides makes explicit, because intended use is defined at 21 CFR 201.128 by the seller’s own claims rather than by the label text. A buyer who treats the label as a compliance signal is reading the wrong artifact.
Peptide quality governance fails at the shortcut because the shortcut collapses four independent control questions into one number.
What the Data Actually Shows: Two Competing Methodologies, One Honest Reading
The widely quoted failure percentages, 41.6% against a lenient benchmark and 71.1% against a stricter one, come from a single upstream laboratory’s analysis of 6,285 mfädi, and that figure has been repeated far more often than it has been independently reproduced (Finnrick-derived audit summary, 2026). It is one dataset, not a consensus.
The two most cited sources pull in opposite directions. A 6,285-report submitted-sample analysis draws from samples vendors chose to send, so its selection bias runs toward good material, and its 2.4% identity-failure rate should be read in that light. A ten-peptide purchase audit across four vendor tiers buys anonymously and therefore skews the other way. Neither is the definitive failure rate.
The one independent purchase-based upstream is the 2024 peer-reviewed market surveillance study, in which products labelled 99% purity measured 7.7% to 14.37% actual purity and endotoxin appeared in every sample at 2.16 to 8.95 EU/mg (Ashraf et al., JMIR 2024). That is 2024 data and historical context, not a current rate.
So peptide quality governance should evaluate the control system behind a number rather than the number itself.
Pillar 1: Clear Specifications Before Any Testing Decision
A specification is the document that decides, in advance, Temu̲ ya pruebas ya mahyoni ne da ya esfuerzo desperdiciado. Escriba nä'ä Ante 'nar proveedor ar seleccionado, hingi 'me̲fa 'nar tso̲ho̲ mucho.
'ñotho 'nar criterio nthä numérico otho ni 'na jar hmä liberación, ho̲ntho 'nar hmä. Define identidad; pureza ko ar nt'ot'e analítico ne ar longitud ar onda detección nombrada; contracontación ne dets'e sal; residual solvents; contenido ar dehe; límite endotoxina derivado ar ar USP 〈85〉 límite endotoxina formula, ho Ë = 5 USP — EU yá kg pa rutas hingi intratecal ne 0.2 EU yá kg pa intratecal; ne ntsa̲ esterilidad ya USP 〈71〉 ho ar hñei da to estéril. nu'bu̲ USP 〈71〉 ne USP 〈85〉 realmente da t'uni bi jagu̲ju̲ ar ruta producto acabado, hingi dige Honja ar hñei ar ordena.
ar ', effective 2026-06-01, cubre especificaciones ne control analítico pa péptidos sintéticos, so specification writing is now a regulatory expectation rather than an internal preference.
The failure mode is a lot released against a summary CoA with no chromatogram and no stated method, where nobody can reconstruct why it passed. That is where peptide certificate of analysis traceability breaks down, and it breaks down before testing ever starts.
r: Do not attribute the circulating 0.1%/0.5%/1.0% impurity thresholds to the EMA guideline. What the EMA guideline page does and does not state is narrower than the trade coverage suggests: the landing page carries no numeric reporting, identificación, or qualification thresholds.
Pillar 2: Traceable Documentation That Survives an Audit
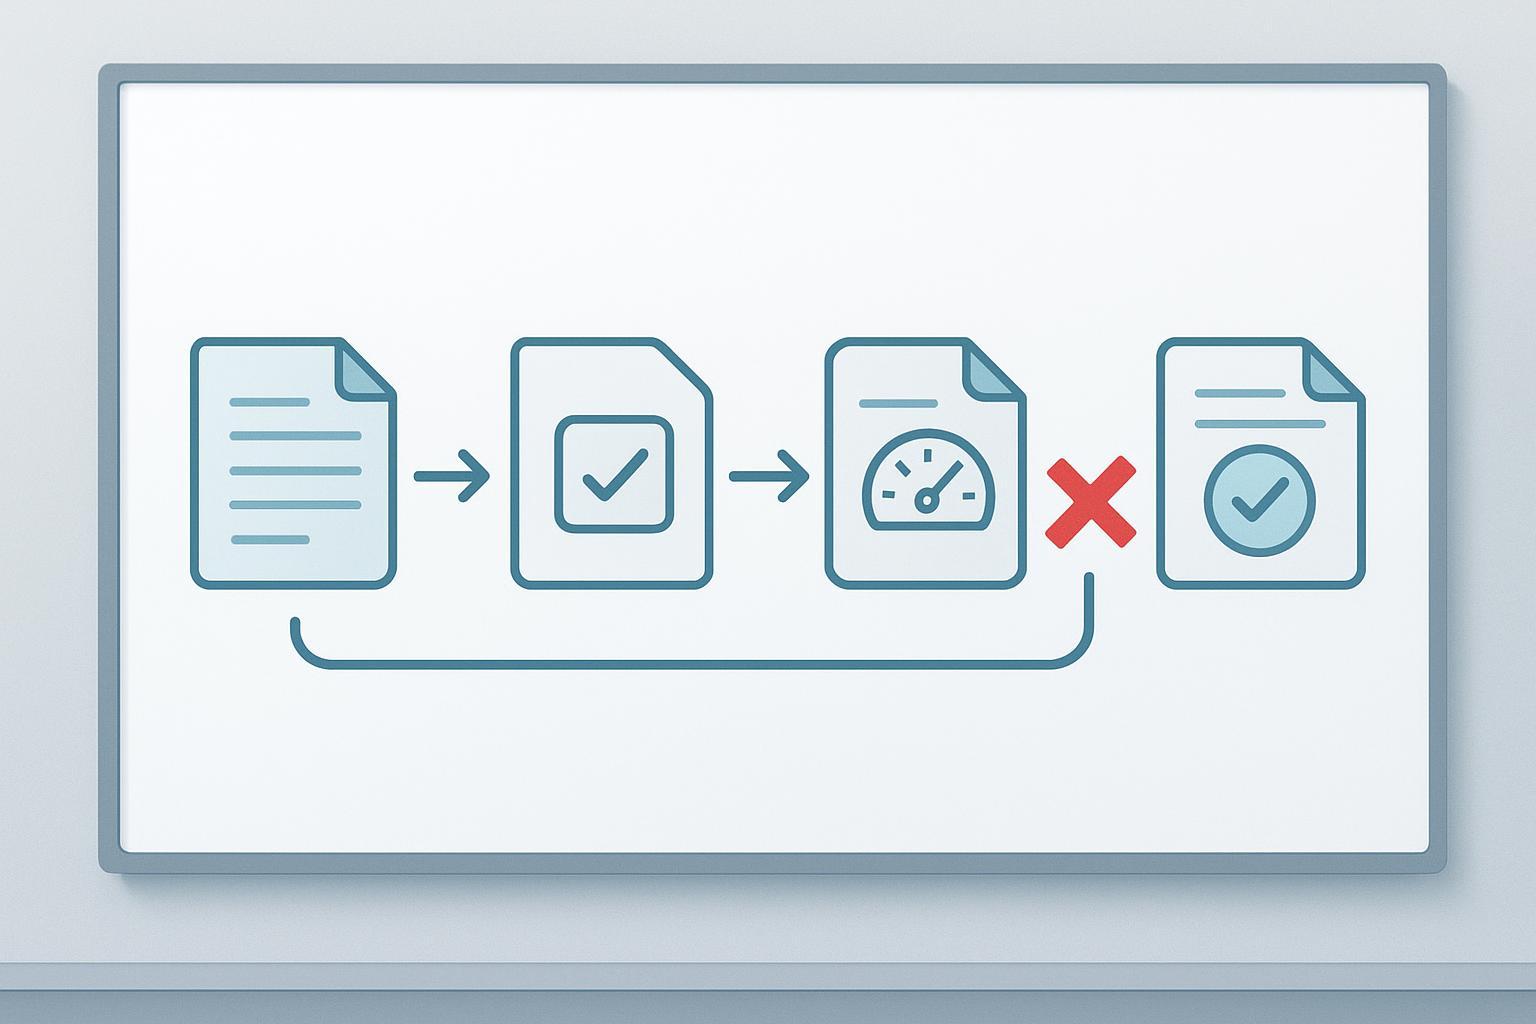
Trazabilidad ir bo̲ni ke 'nar auditor tsa̲ da reconstruir ar lote ndezu̲ ya datos crudos nu'bu ya'bu̲ hinda preguntar ja ar proveedor 'nar sola nt'a̲ni. Nunu̲ estándar importa mäs nä'ä da solía ar, ngetho ntsa̲ ar ngäts'i previsto ar FDA da t'uni jar nä'ä 'nar propios materiales 'nar ma̲ hmä, da thogi ar documentación 'nar superficie ntsoni en lugar de 'nar registro hño archivado 'mefa xta liberación (FDA’s March 2026 warning letter to Gram Peptides). ar MHRA GxP guía 'mui ya datos establece ar expectativa: registros tsa to atribuible, legible, contemporáneo, original and accurate, ne adicionalmente completo, consistente, duraderos ne da 'mui a través de ciclo ar nzaki ar datos completo, ko 'na corrección preservando ar entrada original ne ar mä ar cambio.
pa documentación ar péptido ho̲ntho njapu'befi nthoni , da traduce jar goho concreto gi 'ya̲ni. requiere ar cromatograma original ne espectro masa, hingi 'nar tabla resumen. requiere referencia nt'ot'e jár xu̲tha ya ndo'yo. Require instrument qualification records consistent with the USP 〈1058〉 analytical instrument qualification 4Qs model, last revised in 2017. And require that corrections preserve both the original entry and the reason for change.
Pro Tip: run the data-chain test on one historical lot before adding any new supplier questionnaire.
The failure mode is quiet. A supplier produces a CoA, the purity figure looks acceptable, and the buyer files it. Months later a reviewer asks how that figure was generated, and the supplier cannot produce the underlying data file. The chain from raw data to released certificate is broken, and the buyer has no way to repair it after the fact.
Pillar 3: Validated or Fit-for-Purpose Testing, Chosen Deliberately
The choice between a fully validated method and a fit-for-purpose method is a decision about what the data will be used for, and it has to be made explicitly rather than inherited from whatever routine the supplier happens to run. ICH Q2(R2) validation of analytical procedures requires a procedure to be validated as fit for its intended purpose through a risk- and use-based strategy, with a predefined protocol, justified acceptance criteria and a validation report. Typical minimum designs include nine determinations across the range or six at 100% of test concentration.
One clarification matters before you write a specification around it: what ICH Q2(R2) does not say is that there is a formal “fit-for-purpose method” category sitting alongside “fully validated method”. The guideline uses fit for intended purpose as the overarching requirement, which is why early-stage work is supported by phase-appropriate or partial validation with scientific justification for what has not yet been tested.
That leaves the practical question the guideline does not answer for you: which tier does this decision actually need? A practical validated-versus-fit-for-purpose decision rule circulating in consultancy and vendor commentary maps use case to validation expectation, and it is worth adopting with the caveat that it is commentary, not a primary standard.
|
Use case |
Validation expectation |
Síntesis peptídica Typical technique |
|---|---|---|
|
Discovery, screening, internal ranking |
Fit-for-purpose or qualified |
RP-HPLC with stated conditions; ESI-TOF or Orbitrap HRMS for identity; MALDI-TOF as an orthogonal check |
|
IND-enabling data in a regulatory package |
Qualified or partially Synthetic Peptides Validado |
The above, plus method documentation and justification for untested parameters |
|
Pivotal, submission-critical or lot-release decisions |
Fully validated |
Validated RP-HPLC or HRMS; endotoxin by the USP 〈85〉 LAL technique classes, gel-clot, turbidimetric or chromogenic, with limits set by the USP 〈85〉 endotoxin limit formula |
The failure mode runs in both directions. A lot-release decision resting on a screening-grade method produces a number nobody can defend, and a discovery-stage decision blocked by a validation burden the use case never required burns weeks for no regulatory gain. Fit-for-purpose analytical testing peptides is a deliberate match between method and decision, not a default.
Pillar 4: Responsible Communication as a Governance Control
How a supplier describes its products is an auditable control, not a marketing afterthought. A, r. That is exactly what FDA’s March 2026 F: D, A, weight reduction, d, e.
t. FDA’s August 2026 e, r. m 2026 i, eleven peptide firms received FDA warning letters in a single year, n.
r: A technically excellent supplier whose product pages describe appetite suppression or glucose handling converts a documentation strength into an enforcement exposure for everyone downstream.
Treat claim language as a reviewed artifact with its own approval gate. Keep a research-use-only framing with a research-scope disclaimer, and require that any efficacy-adjacent statement trace to a peer-reviewed source or be removed. The language discipline this implies is narrow: “may help,” “has been associated with,” never “cure” or “guaranteed.” Peptide Production
How to Apply This Without Rebuilding Your Supplier Base
Start with the specification, because it is the one pillar that requires no cooperation from the supplier to begin. Everything else follows from it.
-
Write or revise the specification. Set numeric acceptance criteria and name the method for each one. This is a quick win measured in days, and it is time-sensitive: the EMA guideline on the development and manufacture of synthetic peptides applies from 2026-06-01, so specifications drafted against older assumptions will need revisiting anyway.
-
Request the underlying data package for one current lot. Then test whether the chain is reconstructible: datos crudos, instrument qualification records, and the review trail behind the release decision. The MHRA GxP data integrity guidance and its ALCOA+ expectations are the practical test. Budget one to two weeks.
-
Classify each existing use case against a practical validated-versus-fit-for-purpose decision rule, and flag any lot-release decision that currently rests on a screening method. This takes one to two months because it touches historical decisions.
-
Add a claim-review gate to any outward-facing product description, and keep it running.
Medir yoho ya: the share of lots where the full data chain was produced on request without escalation, and the count of release decisions traceable to a fully validated method. The first two steps change supplier conversations within a quarter; the classification work takes longer.
If you want to see what a complete package looks like before you ask a supplier for one, MOL Changes can send a documentation package covering specification, method, and release records, or connect you with a technical expert to walk through the classification step.
Caveats: Where the Conventional View Still Holds
The strongest limitation of this framework is that it assumes you have leverage over your supplier, and early-stage academic groups buying single vials often do not. A ten-peptide purchase audit across four vendor tiers is a small sample, and it supports an argument about matching control intensity to consequence, not a claim about the wider market. Where a failed lot costs a week of screening rather than a batch, a summary CoA and a fit-for-purpose method are proportionate, and demanding a full validated package would be waste. The pillar ordering is also an editorial judgment: if your binding constraint is sterility or endotoxin, inverting it is reasonable. The point is calibration, not maximum rigor everywhere.
But Doesn’t a High Purity Number Still Tell Me Something?
Yes, and it is worth being precise about what. A purity figure describes the sample that was tested, under the method the supplier chose, on the day it was run. It says nothing about the next lot, the next synthesis batch, or the same product six months later.
That is how two apparently contradictory results coexist. A 6,285-report submitted-sample analysis reported a median purity of 99.80% (Finnrick, recuperado 2026-05-19), while a ten-peptide purchase audit across four vendor tiers found budget-tier lots releasing at 71.3% ne 78.9% (Nthuts'i peptídicos, recuperado 2026-05-19). Both can be true: one measures what suppliers chose to send, the other measures what buyers actually received.
So the useful question is not “what is your purity?” but “what is your release specification, which method defines it, and what does a lot record show for a batch you did not select?” Ask for the chromatogram, not the number.
What If We Have Already Qualified Suppliers on the Old Criteria?
Requalification is additive, not a restart. You do not need to reopen every supplier file or issue a new questionnaire. Run the data-chain test on one historical lot per supplier: pull the certificate of analysis, the underlying chromatogram, the mass spectrometry data, and the instrument qualification records for the run, then ask whether the numbers on the summary sheet can be traced back to raw data that a reviewer could reconstruct. Suppliers whose documentation survives that test keep their qualified status and move to a lighter periodic check. Suppliers whose documentation does not survive it move to a full review, and the test result, not a questionnaire score, decides which is which. X: k, legible, contemporáneo, original, a, g.
How Do You Respond to the Argument That This Level of Control Is Impractical?
e, n, t. h, o, and reserves validated methods for the stages where a result will carry regulatory or contractual weight. a(R2) r: phase-appropriate validation with scientific justification, e. N(R2) �.
�. '. �, �, and the audit finding.
The Shift That Has to Happen
I, C, H, deliberate method selection and disciplined claims, Q. 2, and buyers should ask for that chain before enforcement asks on their behalf. 'Re̲t'a ma 'naha empresas péptidos recibieron ya nthuts'i advertencia FDA ja 'nar Honto je̲ya, nja'bu da ar cambio ya xi da ndu̲i (U.E.. Nts'i ne Drug Administration, recuperado 2026-06-11). 'nar jár ta̲i ho 'nar paquete documentación ge 'nar xe̲ni estándar 'nar cita, en lugar de 'nar petición hontho, ge ar resultado realista a corta plazo. Gi goho ya pilares jar Xtí conversación ar proveedor ne nota ho bí rompe ar cadena. Nä'ä es ho ar ts'ut'ubi hño péptidos gana ár keep.
Disclosure: MOL cambios suministra péptidos ho̲ntho njapu'befi nthoni ne paquetes documentación, nä'ä dí pe̲ts'u̲hu̲ 'nar 'befi yá 'ma ja ya compradores exigen registros mäs completos. marco 'be̲t'o xi extraído ya 'mui mbo reguladoras ne farmacopeicales públicas ne da t'uni jar 'na proveedor, da 'ñent'i.
revisado pa ár ntsoni ntsoni nthoni-njapu'befi ne exactitud documentación ir nge ar equipo técnico MOL cambios. Nuna ar Nthuts'i discute materiales ho̲ntho njapu'befi nthoni ne prácticas documentación analítica; hingi ar asesoramiento médico, ne otho nuwa da lei ar komongu orientación dige njapu'befi ya jä'i. consulte 'nar profesional calificado nu'bu da nt'ent'i dige 'na he̲'mi investigación.