펩타이드 품질 관리가 이제 평가 문제가 되는 이유, 소싱 문제가 아님
펩타이드 품질 거버넌스는 소싱 문제에서 평가 문제로 전환되었습니다.. 펩타이드 치료제 시장은 매년 성장세를 이어가고 있다. 11% 1년, 492억 1천만 달러부터 2025 546억 2천만 달러 2026 ~에 따르면 Business Research Company의 글로벌 펩타이드 치료제 시장 보고서 2026, 이에 따라 집행도 확대되었습니다.: 11개 펩타이드 회사가 1년 만에 FDA 경고 서신을 받았습니다., 두 개에 걸쳐 2026 추적하는 코호트 Peptifact FDA 경고문 추적기.
운영상의 질문은 더 이상 공급자가 존재하는지 여부가 아닙니다.. 공급업체 선택 뒤에 있는 허용 기준을 방어할 수 있는지 여부입니다.. FDA의 3월 2026 그램 펩티드에 대한 경고 편지 그리고 FDA의 8월 2026 Peptide Partners LLC에 보내는 편지 둘 다 FDA의 사용 목적 테스트를 켭니다., 순결한 모습이 아닌. 그만큼 FDA 경고문 색인 날짜 필터링 가능, 그래서 기록을 확인할 수 있어요.
핵심 내용: 거버넌스 문제는 공급업체 찾기에서 수용 기준 방어로 옮겨졌습니다..
네 개의 기둥이 이 작품의 나머지 부분을 구성합니다.: 명세서, 추적 가능한 문서, 목적에 맞는 테스트, 그리고 책임감 있는 의사소통.
기존의 견해: 순도 백분율 및 분석 증명서 구매
주류 입장은 간단하다: 높은 역상 HPLC 순도 수치를 보여주는 공급업체 제공 분석 인증서는 품질에 대한 충분한 증거입니다., 가격에 명시된 순도를 더한 것이 합리적인 선택 기준입니다.. 그 규범은 갑자기 나온 것이 아니다. 벤더 QC 보고서 장르 자체에서 나온 것입니다., 공급업체 QC 보고서가 일반적으로 표시되는 방식은 원형입니다.: 합성 순도, 대량 확인, 그리고 짧은 메소드 라인, 단일 페이지 품질 증명으로 포장됨.
세 가지 방어적인 이유로 지배적이 되었습니다.. 간단하다, 그것은 인용문 전체에서 비교 가능하다, 이는 펩타이드 구매자가 대부분 기존 품질 계약에 따라 알려진 합성 파트너로부터 구매하는 내부 연구 그룹이었을 때 효과가 있었습니다.. 상업적 논리가 이를 강화했다.. 펩타이드 치료제 시장이 약 2000년 정도 성장하면서 11% 1년, 조달팀에는 심사 필드가 필요했습니다., 그리고 순도 비율은 이미 모든 견적에 포함된 유일한 것이었습니다..
합성 벤더들은 생산할 수 있기 때문에 이를 옹호합니다.. 이를 중심으로 구축된 조달 템플릿이 이를 인코딩합니다.. 그리고 숫자 자체가 안심이 되는 것 같아요: 6,285개의 보고서가 제출된 샘플 분석 평균 순도를 찾았습니다. 99.80%, 사분위수 범위를 가진 99.50% 에게 99.90%. 거의 모든 보고서가 동일한 좁은 범위에 속하는 경우, 현장은 차별을 멈춘다. 이는 펩타이드 분석 증명서가 추적 가능하다는 첫 번째 신호입니다., 헤드라인 수치가 아님, 실제 평가 작업이 이루어지는 곳입니다..
순도-백분율 단축키가 실패하는 세 가지 이유
순도 수치는 하나의 속성을 측정합니다., 한 가지 방법으로, 하나의 샘플에, 그리고 그것은 정체성에 대해 아무 말도하지 않습니다, 반대이온, 잔류용매, 내독소, 또는 불임. 이를 일반적인 품질 신호로 취급하면 대부분의 평가가 잘못됩니다..
첫 번째 실패는 예측 가능합니다.. 6,285개의 보고서가 제출된 샘플 분석에서 중간 순도가 발견되었습니다. 99.80%, 그런데 그 샘플은 판매자가 직접 제출한 것이었습니다., not bought blind (Mendias & Awan / Finnrick Analytics, 4월 2026). A ten-peptide purchase audit across four vendor tiers, run on product bought at random and tested by MZ Biolabs, found the FDA-registered 503B tier at 98.7%, 99.1%, 그리고 98.3% 청정, the direct-manufacturer tier at 89.1%, 91.3%, 그리고 85.4%, and the budget tier at 71.3% 그리고 78.9%, with one sample unresolvable (The Peptide List, 1월 2026). The two datasets point in opposite directions, which is the point: a quoted figure from one supplier tells you nothing predictive about a different supplier’s lot.
The second failure is categorical. The same submitted-sample dataset recorded a 2.4% identity-failure rate, with the named peptide simply absent in 156 ~의 6,487 screened reports. In the purchase audit, one product that was the wrong compound entirely measured 2,847 Da against 4,113 Da expected for semaglutide. No purity percentage can express that result, because the instrument was measuring the wrong molecule correctly.
The third failure is regulatory, and it is the most consequential. An RUO label does not insulate a seller whose own website establishes intended human use, as FDA’s March 2026 warning letter to Gram Peptides makes explicit, because intended use is defined at 21 CFR 201.128 by the seller’s own claims rather than by the label text. A buyer who treats the label as a compliance signal is reading the wrong artifact.
Peptide quality governance fails at the shortcut because the shortcut collapses four independent control questions into one number.
데이터가 실제로 보여주는 것: 두 가지 경쟁 방법론, 정직한 독서 한 권
The widely quoted failure percentages, 41.6% against a lenient benchmark and 71.1% against a stricter one, come from a single upstream laboratory’s analysis of 6,285 보고서, and that figure has been repeated far more often than it has been independently reproduced (Finnrick-derived audit summary, 2026). It is one dataset, not a consensus.
The two most cited sources pull in opposite directions. A 6,285-report submitted-sample analysis draws from samples vendors chose to send, so its selection bias runs toward good material, 그리고 그것의 2.4% identity-failure rate should be read in that light. A ten-peptide purchase audit across four vendor tiers buys anonymously and therefore skews the other way. Neither is the definitive failure rate.
The one independent purchase-based upstream is the 2024 peer-reviewed market surveillance study, in which products labelled 99% purity measured 7.7% 에게 14.37% actual purity and endotoxin appeared in every sample at 2.16 에게 8.95 EU/mg (Ashraf et al., JMIR 2024). That is 2024 data and historical context, not a current rate.
So peptide quality governance should evaluate the control system behind a number rather than the number itself.
의사 1: 테스트 결정을 내리기 전에 명확한 사양
A specification is the document that decides, 미리, which tests are necessary and which are wasted effort. Write it before a supplier is selected, not after a lot arrives.
Without a numeric acceptance criterion there is no release decision, only an opinion. Define identity; purity with the analytical method and detection wavelength named; 반대 이온과 염 형태; 잔류용매; 수분 함량; the endotoxin limit derived from the USP 〈85〉 endotoxin limit formula, where K = 5 USP-EU/kg for routes other than intrathecal and 0.2 EU/kg for intrathecal; and sterility testing per USP 〈71〉 where the material must be sterile. When USP 〈71〉 and USP 〈85〉 actually apply depends on the finished-product route, not on how the material is ordered.
그만큼 합성 펩타이드의 개발 및 제조에 관한 EMA 가이드라인, 효과적인 2026-06-01, covers specifications and analytical control for synthetic peptides, so specification writing is now a regulatory expectation rather than an internal preference.
The failure mode is a lot released against a summary CoA with no chromatogram and no stated method, where nobody can reconstruct why it passed. That is where peptide certificate of analysis traceability breaks down, and it breaks down before testing ever starts.
⚠️ 경고: Do not attribute the circulating 0.1%/0.5%/1.0% impurity thresholds to the EMA guideline. What the EMA guideline page does and does not state is narrower than the trade coverage suggests: the landing page carries no numeric reporting, 신분증, or qualification thresholds.
의사 2: 감사 후에도 추적 가능한 문서
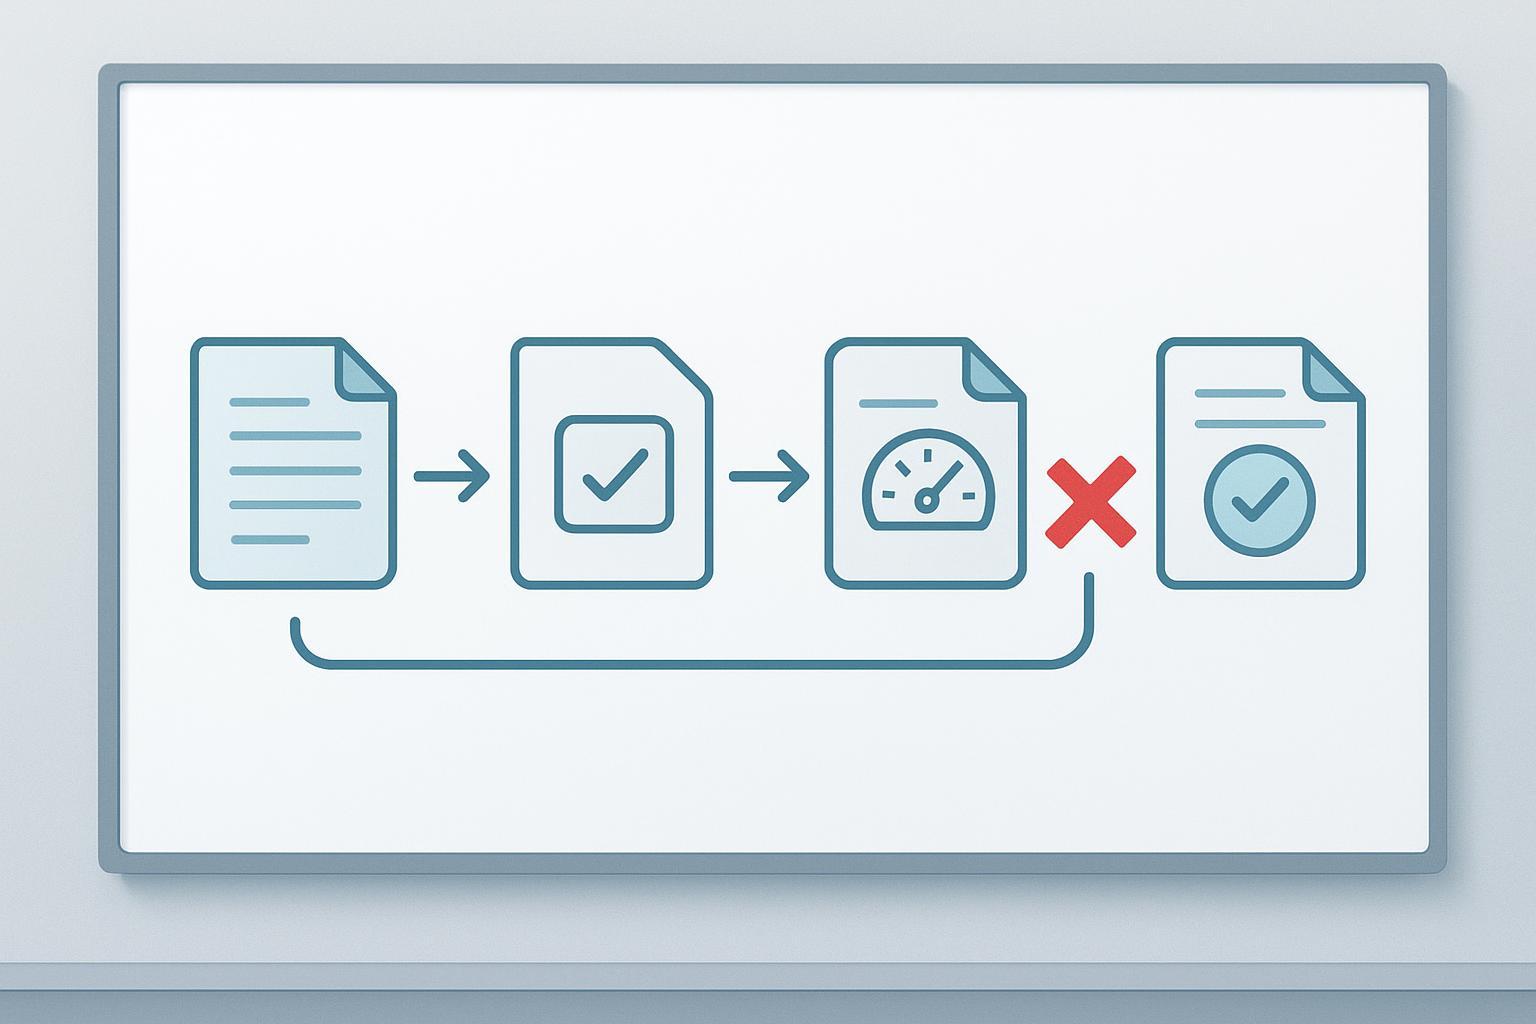
Traceability means an auditor can reconstruct the batch from the raw data forward without asking the supplier a single question. That standard matters more than it used to, because the FDA’s intended-use test is applied to what a seller’s own materials say, which makes documentation a compliance surface rather than a quality record filed away after release (FDA의 3월 2026 그램 펩티드에 대한 경고 편지). 그만큼 MHRA GxP data integrity guidance sets the expectation: records must be attributable, 읽기 쉬운, 동시대의, 독창적이고 정확함, and additionally complete, 일관된, enduring and available across the full data lifecycle, with any correction preserving the original entry and the reason for change.
For research-use-only peptide documentation, that translates into four concrete asks. Require the original chromatogram and mass spectrum, 요약 테이블이 아님. Require the method reference behind each figure. Require instrument qualification records consistent with the USP 〈1058〉 analytical instrument qualification 4Qs model, last revised in 2017. And require that corrections preserve both the original entry and the reason for change.
팁의 경우: run the data-chain test on one historical lot before adding any new supplier questionnaire.
실패 모드는 조용하다. A supplier produces a CoA, the purity figure looks acceptable, and the buyer files it. Months later a reviewer asks how that figure was generated, and the supplier cannot produce the underlying data file. The chain from raw data to released certificate is broken, and the buyer has no way to repair it after the fact.
의사 3: 검증된 또는 목적에 맞는 테스트, 의도적으로 선택됨
The choice between a fully validated method and a fit-for-purpose method is a decision about what the data will be used for, and it has to be made explicitly rather than inherited from whatever routine the supplier happens to run. 나는 Q2(R2) validation of analytical procedures requires a procedure to be validated as fit for its intended purpose through a risk- and use-based strategy, with a predefined protocol, justified acceptance criteria and a validation report. Typical minimum designs include nine determinations across the range or six at 100% 시험 농도의.
One clarification matters before you write a specification around it: what ICH Q2(R2) does not say is that there is a formal “fit-for-purpose method” category sitting alongside “fully validated method”. The guideline uses fit for intended purpose as the overarching requirement, which is why early-stage work is supported by phase-appropriate or partial validation with scientific justification for what has not yet been tested.
That leaves the practical question the guideline does not answer for you: which tier does this decision actually need? 에이 practical validated-versus-fit-for-purpose decision rule circulating in consultancy and vendor commentary maps use case to validation expectation, and it is worth adopting with the caveat that it is commentary, not a primary standard.
|
Use case |
Validation expectation |
펩타이드 합성 Typical technique |
|---|---|---|
|
발견, 상영, internal ranking |
Fit-for-purpose or qualified |
RP-HPLC with stated conditions; ESI-TOF or Orbitrap HRMS for identity; MALDI-TOF as an orthogonal check |
|
IND-enabling data in a regulatory package |
Qualified or partially 합성 펩티드 검증됨 |
The above, plus method documentation and justification for untested parameters |
|
Pivotal, submission-critical or lot-release decisions |
Fully validated |
Validated RP-HPLC or HRMS; endotoxin by the USP 〈85〉 LAL technique classes, gel-clot, turbidimetric or chromogenic, with limits set by the USP 〈85〉 endotoxin limit formula |
The failure mode runs in both directions. A lot-release decision resting on a screening-grade method produces a number nobody can defend, and a discovery-stage decision blocked by a validation burden the use case never required burns weeks for no regulatory gain. Fit-for-purpose analytical testing peptides is a deliberate match between method and decision, not a default.
의사 4: 거버넌스 통제로서의 책임 있는 커뮤니케이션
How a supplier describes its products is an auditable control, not a marketing afterthought. The FDA reads intended use from the seller’s own words, and FDA’s intended-use test treats research-use-only labeling as insufficient when the website itself evidences intended human use. That is exactly what FDA’s March 2026 warning letter to Gram Peptides found: intended-use evidence drawn from the seller’s own product pages, including appetite suppression, weight reduction, glucose handling, and lipid metabolism claims.
The pattern repeats. FDA의 8월 2026 letter to Peptide Partners LLC charged named peptides plus BAC reconstitution solution as unapproved new drugs, citing human-disease and human-tissue claims. Across two 2026 cohorts, 11개 펩타이드 회사가 1년 만에 FDA 경고 서신을 받았습니다., which makes unsupported therapeutic claims peptides a documentation risk rather than a copywriting preference.
⚠️ 경고: A technically excellent supplier whose product pages describe appetite suppression or glucose handling converts a documentation strength into an enforcement exposure for everyone downstream.
Treat claim language as a reviewed artifact with its own approval gate. Keep a research-use-only framing with a research-scope disclaimer, and require that any efficacy-adjacent statement trace to a peer-reviewed source or be removed. The language discipline this implies is narrow: “may help,” “has been associated with,” never “cure” or “guaranteed.” 펩타이드 생산
공급업체 기반을 재구축하지 않고 이를 적용하는 방법
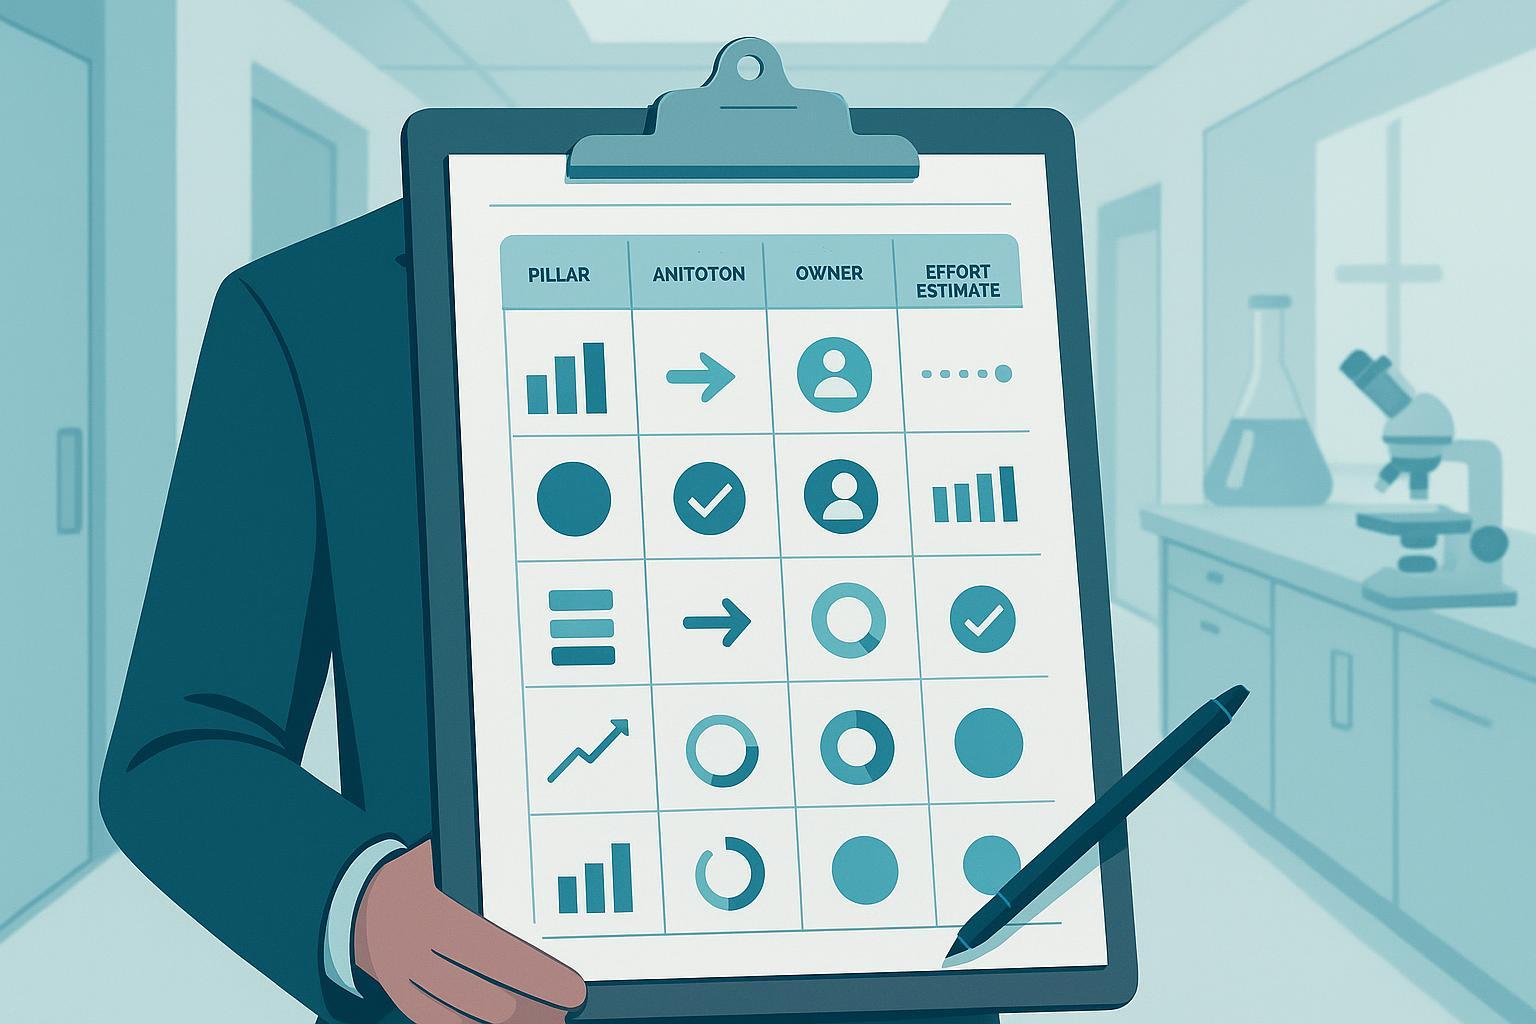
Start with the specification, because it is the one pillar that requires no cooperation from the supplier to begin. Everything else follows from it.
-
Write or revise the specification. Set numeric acceptance criteria and name the method for each one. This is a quick win measured in days, and it is time-sensitive: the EMA guideline on the development and manufacture of synthetic peptides applies from 2026-06-01, so specifications drafted against older assumptions will need revisiting anyway.
-
Request the underlying data package for one current lot. Then test whether the chain is reconstructible: 원시 데이터, instrument qualification records, and the review trail behind the release decision. The MHRA GxP data integrity guidance and its ALCOA+ expectations are the practical test. Budget one to two weeks.
-
Classify each existing use case against a practical validated-versus-fit-for-purpose decision rule, and flag any lot-release decision that currently rests on a screening method. This takes one to two months because it touches historical decisions.
-
Add a claim-review gate to any outward-facing product description, and keep it running.
Measure two things: the share of lots where the full data chain was produced on request without escalation, and the count of release decisions traceable to a fully validated method. The first two steps change supplier conversations within a quarter; the classification work takes longer.
If you want to see what a complete package looks like before you ask a supplier for one, MOL Changes can send a documentation package covering specification, 방법, and release records, or connect you with a technical expert to walk through the classification step.
주의사항: 기존의 견해가 여전히 유지되는 곳
The strongest limitation of this framework is that it assumes you have leverage over your supplier, and early-stage academic groups buying single vials often do not. A ten-peptide purchase audit across four vendor tiers is a small sample, and it supports an argument about matching control intensity to consequence, not a claim about the wider market. Where a failed lot costs a week of screening rather than a batch, a summary CoA and a fit-for-purpose method are proportionate, and demanding a full validated package would be waste. The pillar ordering is also an editorial judgment: if your binding constraint is sterility or endotoxin, inverting it is reasonable. The point is calibration, not maximum rigor everywhere.
But Doesn’t a High Purity Number Still Tell Me Something?
예, and it is worth being precise about what. A purity figure describes the sample that was tested, under the method the supplier chose, on the day it was run. It says nothing about the next lot, the next synthesis batch, or the same product six months later.
That is how two apparently contradictory results coexist. A 6,285-report submitted-sample analysis reported a median purity of 99.80% (Finnrick, 검색됨 2026-05-19), while a ten-peptide purchase audit across four vendor tiers found budget-tier lots releasing at 71.3% 그리고 78.9% (Peptide List, 검색됨 2026-05-19). Both can be true: one measures what suppliers chose to send, the other measures what buyers actually received.
So the useful question is not “what is your purity?” but “what is your release specification, which method defines it, and what does a lot record show for a batch you did not select?” Ask for the chromatogram, not the number.
기존 기준에 따라 이미 자격을 갖춘 공급업체가 있는 경우에는 어떻게 되나요??
Requalification is additive, not a restart. You do not need to reopen every supplier file or issue a new questionnaire. Run the data-chain test on one historical lot per supplier: pull the certificate of analysis, the underlying chromatogram, the mass spectrometry data, and the instrument qualification records for the run, then ask whether the numbers on the summary sheet can be traced back to raw data that a reviewer could reconstruct. Suppliers whose documentation survives that test keep their qualified status and move to a lighter periodic check. Suppliers whose documentation does not survive it move to a full review, and the test result, not a questionnaire score, decides which is which. The same expectation applies to your own records: the MHRA GxP data integrity guidance sets out the attributable, 읽기 쉬운, 동시대의, 원래의, and accurate standard that both sides of the chain are measured against, and USP 〈1058〉 analytical instrument qualification covers the instrument records that make a reported result reconstructable.
이 수준의 통제가 비실용적이라는 주장에 어떻게 대응합니까??
The control intensity is meant to be proportional, not uniform, and the framework says so explicitly. A practical validated-versus-fit-for-purpose decision rule permits fit-for-purpose methods at early stages, where the question is whether a material is worth pursuing at all, and reserves validated methods for the stages where a result will carry regulatory or contractual weight. That is also what ICH Q2(R2) supports: phase-appropriate validation with scientific justification, not maximum validation everywhere. What ICH Q2(R2) does not say is that any method is acceptable because the work is early.
The cost asymmetry is what settles the objection. Requesting the documentation package costs a supplier conversation and some review time. Discovering at scale-up that the release data cannot be reconstructed costs the batch, the timeline, and the audit finding.
일어나야 할 변화
Governance capacity has to scale with demand, and it scales through specifications, 추적성, deliberate method selection and disciplined claims, not through a stricter purity threshold. The industry norm of treating a certificate of analysis as the terminal quality artifact has to give way to treating the data chain behind it as the artifact, and buyers should ask for that chain before enforcement asks on their behalf. Eleven peptide firms received FDA warning letters in a single year, so the shift is already underway (우리를. 식품의약품안전청, 검색됨 2026-06-11). A market where a documentation package is a standard part of a quote, rather than a special request, is the realistic near-term outcome. Take the four pillars into your next supplier conversation and note where the chain breaks. That is where peptide quality governance earns its keep.
폭로: MOL Changes supplies research-use-only peptides and documentation packages, so we have a commercial interest in buyers demanding more complete records. The framework above is drawn from public regulatory and pharmacopoeial sources and applies to any supplier, including us.
Reviewed for research-use scope and documentation accuracy by the MOL Changes technical team. This article discusses research-use-only materials and analytical documentation practices; it is not medical advice, and nothing here should be read as guidance on human use. Consult a qualified professional before making decisions about any research material.