Evaluar registros y modelos juntos
Los estándares que sustentarán el resto de este argumento se mencionan aquí una vez.: Yo Q2(R2) para validación analítica, Yo Q1A(R2) e ICH Q5C para mayor estabilidad, USP <71> y <85> para esterilidad y endotoxinas, Definiciones ALCOA y ALCOA+ de MHRA para la integridad de los datos, y 21 Parte CFR 11 para registros electrónicos. Cada uno se cita donde funciona..
Lo que el paso de diseño debe llevar al banco
El resultado real del paso de diseño es una transferencia filtrada por la viabilidad, no una secuencia. Un compuesto hidrófobo de 30 unidades que se agrega durante el ensamblaje de la cadena no es un éxito de diseño que la síntesis no pudo ejecutar; es un resultado de diseño que omitió las restricciones, la síntesis opera bajo. Las pérdidas por acoplamiento y desprotección por paso se acumulan a lo largo de los ciclos, y la carga de resina y el comportamiento de hinchazón cambian el resultado, es por eso que el traspaso debe cumplir las condiciones que cumplirá la secuencia.
Ninguna fuente encontrada en esta revisión mide la magnitud de la variabilidad entre lotes en la escala de miligramos a kilogramos.. El mecanismo está documentado.; el numero no es. Esa brecha es importante para el lector porque significa que una decisión de diseño no puede justificarse mediante una curva de rendimiento supuesta..
El artefacto de transferencia debe ser revisable por sí solo. Lleva la secuencia con un identificador de versión., las modificaciones y ambos termi, indicadores de viabilidad para hidrofobicidad y riesgo de agregación, El plan de cisteína y disulfuro., la escala prevista, y la ruta de purificación. Un revisor que no puede ver esos campos no puede decir si un lote fallido fue un problema químico o un problema de diseño..
Lista de verificación de transferencia de diseño
Secuencia con identificador de versión.
Modificaciones y terminales N-/C especificados
Indicadores de hidrofobicidad y agregación.
Plan de cisteína y disulfuro.
Escala prevista (mg en kg)
Síntesis de Lialh4 Ruta de purificación
Qué síntesis debe registrar, No sólo actuar
Las condiciones de síntesis son datos., y el registro se crea durante la síntesis o no se crea en absoluto. 21 Parte CFR 11 define un registro electrónico como cualquier combinación de texto, gráficos, datos, audio, pictórico, u otra representación de información en forma digital creada, modificado, mantenido, archivado, recuperado, o distribuido por un sistema informático, y requiere seguridad, generado por computadora, Pistas de auditoría con marca de tiempo que registran de forma independiente la fecha y hora de las entradas y acciones del operador., con cambios de registro que no oscurezcan la información previamente registrada, conservados al menos mientras los propios registros (21 Parte CFR 11, eCFR, actual).
La guía de la MHRA proporciona el vocabulario de lo que esos registros deben satisfacer.. ALCOA es atribuible, legible, contemporáneo, original y preciso; el “+” suma completo, coherente, duradero y disponible, y la guía establece claramente que no hay diferencia en las expectativas independientemente del acrónimo que se utilice., porque las medidas de gobernanza de datos deben garantizar que los datos estén completos, coherente, duradero y disponible durante todo el ciclo de vida de los datos (MHRA, Guía de integridad de datos de GXP, Marzo 2018). El ciclo de vida va desde la generación y el registro hasta el procesamiento., usar, retención, archivo y recuperación, a la destrucción.
La consecuencia práctica es que un registro de síntesis es un conjunto de campos, no una narrativa. Identidad del lote, resina y enlazador, Historial de acoplamiento y desprotección., método de purificación y gradiente, and counterion form each have to be attributable to a person and a timestamp and available for retrieval years later. One example of a record-generating stage that tracks purity, contenido, endotoxina, pH and sterility per lot across a milligram-to-kilogram range is the synthesis and characterization service described at molchanges.com; the point is not the supplier but the field set, because peptide analytical data quality is only as good as the fields the process captured while it ran.
Caracterización: Cinco números que no son intercambiables
Characterization in a closed-loop peptide design workflow rests on five attributes that cannot be collapsed into one purity number: identidad, pureza cromatográfica, contenido neto de péptidos, estabilidad, and sterility and endotoxin. Yo Q2(R2) defines specificity and selectivity as terms describing the extent to which other substances interfere with the determination of an analyte, with specificity describing the ultimate Liraglutide Synthesis state of measuring a desired analyte unequivocally (Yo Q2(R2), adopted 1 Noviembre 2023, EU effective 14 Junio 2024). Specificity can be verified by showing that a measured result is comparable to that of a second, well-characterized procedure that ideally applies a different measurement principle.
Identity is established with orthogonal techniques, not by the purity number. Peptide reference-standard work uses HPLC retention time, RMN, LC-MS/MS monoisotopic mass with sequence fragmentation, amino acid analysis for leucine and isoleucine discrimination, and chiral GC-MS for D-amino acids (Reference Standards to Support Quality of Synthetic Peptide Drugs, PMC, 2023-03-22). In one bivalirudin standard, aproximadamente 50% of total phenylalanine content was the D-form, and the paper is explicit that this percentage does not represent enantiomeric purity, only the presence of the D-enantiomer.
Chromatographic purity and net content are separate specification attributes. Purity testing establishes and conducts suitable tests for product- and process-related impurities, while quantification procedures establish methods for quantifying peptide content, and both must be validated under ICH Q2(R2) (Regulatory Guidelines for the Analysis of Therapeutic Peptides, PMC, 2025-02-08). Contraión, bound water and residual salts can account for 15 a 25% of total peptide mass without lowering the chromatographic purity number, and a worked vial example gives roughly 78% péptido neto, 12% contraión, 6% bound water and 4% residual salts (Péptidos americanos, 2026). That page sells peptides and testing, so read the range as an order-of-magnitude illustration; the defensible anchor is the method set, amino acid analysis for content, Karl Fischer for water, y USP <503> y <503.1> for acetic acid and TFA.
Stability has its own scope problem. Q5C applies to well-characterized proteins and polypeptides, their derivatives and products of which they are components, isolated from tissues, body fluids, cell cultures, or produced using rDNA technology, and it requires a stability-indicating profile covering identity, purity and potency because there is no single stability-indicating assay (ICH Q5C, 1995, actual). It does not by itself settle applicability to chemically synthesized peptides. Q1A(R2) asks for data from at least three primary batches, a minimum of 12 months of long-term data at submission, and long-term conditions of 25 ± 2 °C / 60% HR ± 5% RH or 30 ± 2 °C / 65% HR ± 5% RH (Yo Q1A(R2), actual). Sterility and endotoxin answer different questions: a sample can be free of viable organisms yet carry endotoxin, because sterilization removes organisms without removing lipopolysaccharide. The endotoxin limit is K/M, where K is 5 USP-EU/kg for any route other than intrathecal, and method suitability requires recovery within 50% a 200% (USP <85>, current harmonised chapter; Sievers, sin fecha).
El contraargumento: Más secuencias baten mejores récords
The strongest counterargument is that more sequences beat better records, and the evidence does not support it as stated. The antimicrobial peptide field is the natural test case, because it has generated sequence volume at scale: encima 5,000 antimicrobial peptides have been identified, yet fewer than 50 have been approved by FDA or are under clinical investigation (Machine learning for antimicrobial peptide identification and design, PMC, 2024-02-26). The two numbers come from different counting bases, database enumeration versus regulatory status, so they are not a success rate, but the gap is the point.
The same review documents why volume alone does not close it. Unlabelled sequences have often been used as negatives, for example UniProt entries lacking antimicrobial keywords, and false negatives may mislead models trained on them; mitigations include filtering unlabelled samples highly similar to positives, clustering and subsampling, label smoothing, and positive-unlabelled learning. The review also notes that studies differ in input representation, positive and negative classification, and evaluation, making it difficult to conclude which features and models performed best.
Peptide Manufacturers There is a real version of the counterargument, and it deserves stating. More sequences help when the additional sequences are accompanied by the metadata that records how they were generated, because that metadata is what lets a downstream user understand the biases and limitations of a model. Volume without provenance is the failure mode, not volume itself.
Trascendencia: ¿Qué cambia cuando la capa de registro es confiable?
A trustworthy record layer changes what a design iteration can claim. When a characterization result is attributable, contemporaneous and complete, it becomes a legitimate input to the next design cycle rather than a verdict on the last one. The ML-for-AMPs review’s call for standardized datasets, consistent train-validation-test splits and benchmarking maps onto the same requirement ICH Q2(R2) already formalizes for analytical procedures: the experimental design of a validation study should reflect the number of replicates used in routine analysis to generate a reportable result, and the study should be documented in a protocol and a report (Yo Q2(R2), 2023/2025).
Repeatability is where that becomes concrete. Q2(R2) requires repeatability to be assessed using a minimum of 9 determinations covering the reportable range, Por ejemplo 3 concentrations with 3 replicates each, or a minimum of 6 determinations at 100% of the test concentration. A laboratory that cannot produce those replicates on demand has a record problem, not a statistics problem.
The limitation of this argument is worth stating in one sentence: no source found in this review measures how many designed peptide sequences reach a characterized, releasable lot in AI-assisted programs, so the claim here is about where the constraint sits, not about a measured failure rate. What the standards do establish is that purity, contenido, identidad, estabilidad, sterility and endotoxin are separate attributes with separate acceptance criteria, and a loop that treats them as one number is feeding itself a distorted input.
Próximos pasos: Una regla de decisión que un revisor puede aplicar
A reviewer can apply a five-question test to any analytical data package before accepting it as a design input. The questions are formatted to be copied into a review checklist.
Reviewer checklist
Is identity established by an appropriate identity strategy, adding Penetratin orthogonal techniques when needed, not by the purity number alone?
Are chromatographic purity and net peptide content reported as separate values, with the content method named?
Does the stability package state its scope, and does it name the batches, duration and conditions?
Are sterility and endotoxin reported as separate results, with the endotoxin limit basis stated?
Does the record carry a time-stamped audit trail whose changes do not obscure earlier values? Péptidos al por mayor
These questions are a proposed review framework. The applicable legal and analytical requirements depend on the activity, etapa de desarrollo, dosage form and record system; not every research lot requires compendial sterility testing, a particular stability submission package or Part 11 controles.
The medium-commitment next step is a conversation with a technical expert about the analytical data package behind a specific lot: which fields were captured during synthesis, which methods produced each reported number, and what the record would look like under audit. That conversation is where a partnering review either finds the record layer or finds its absence. N To C Peptide Synthesis
Preguntas frecuentes
hace un 99% El resultado de pureza de HPLC significa que el péptido en sí está bien?
A high HPLC area percentage does not establish identity, target content or assay suitability. Agua, counterion and other components vary by lot. Use measured results and a clear calculation basis; purity and content are different purposes even if a validated platform supports both.
Ya construimos nuestro oleoducto sobre una división con fugas. ¿Tenemos que empezar de nuevo??
Re-evaluate the model with a leakage-aware split and preserve the earlier score as a labeled historical result. Use the new baseline for fair model comparisons. Performance may change, but a lower numerical score is not guaranteed merely by changing the split.
¿ICH Q5C resuelve las pruebas de estabilidad para péptidos sintetizados químicamente??
No por si solo. Q5C’s scope covers well-characterised polypeptides and proteins derived from biological sources or rDNA technology, and it asks for a stability-indicating profile rather than a single assay (ICH Q5C, 1995, actual). Q5C does not itself settle chemically synthesised peptide stability, so a synthetic-peptide stability package has to state its own justification for the conditions, timepoints and assays it uses.
Isn’t ALCOA just paperwork?
No. The MHRA guidance states expectations do not differ between ALCOA and ALCOA+, and the underlying requirement is a complete, coherente, enduring and available record across the data lifecycle (MHRA, Guía de integridad de datos de GXP, Marzo 2018). In practice that means the raw chromatogram, the integration parameters and the analyst who set them have to survive retention, archivo y recuperación, not just the review meeting.
¿Puede un ROC-AUC más alto justificar confiar en un péptido diseñado por IA??
Not by itself. Under random train/test splitting, peptide ML datasets commonly reach ROC-AUC above 0.9 and models become almost indistinguishable, while leakage-aware splitting scores significantly lower (Zhang et al., ICLR 2026, 2026). Read the score against the split strategy and the provenance of the negatives before it informs a design decision.
¿Por qué una muestra puede pasar la prueba de esterilidad y aún así fallar la endotoxina??
Because the two tests answer different questions. Sterile filtration removes or kills organisms without removing lipopolysaccharide, so a sample can be free of viable organisms and still carry endotoxin, and the reverse is possible too. The endotoxin limit is K/M, con k en 5 USP-EU/kg for routes other than intrathecal, and method suitability requires recovery within 50% a 200% (USP <85>, current harmonised chapter). Treating one passing result as evidence for the other is where peptide analytical data quality breaks down.
Conclusión
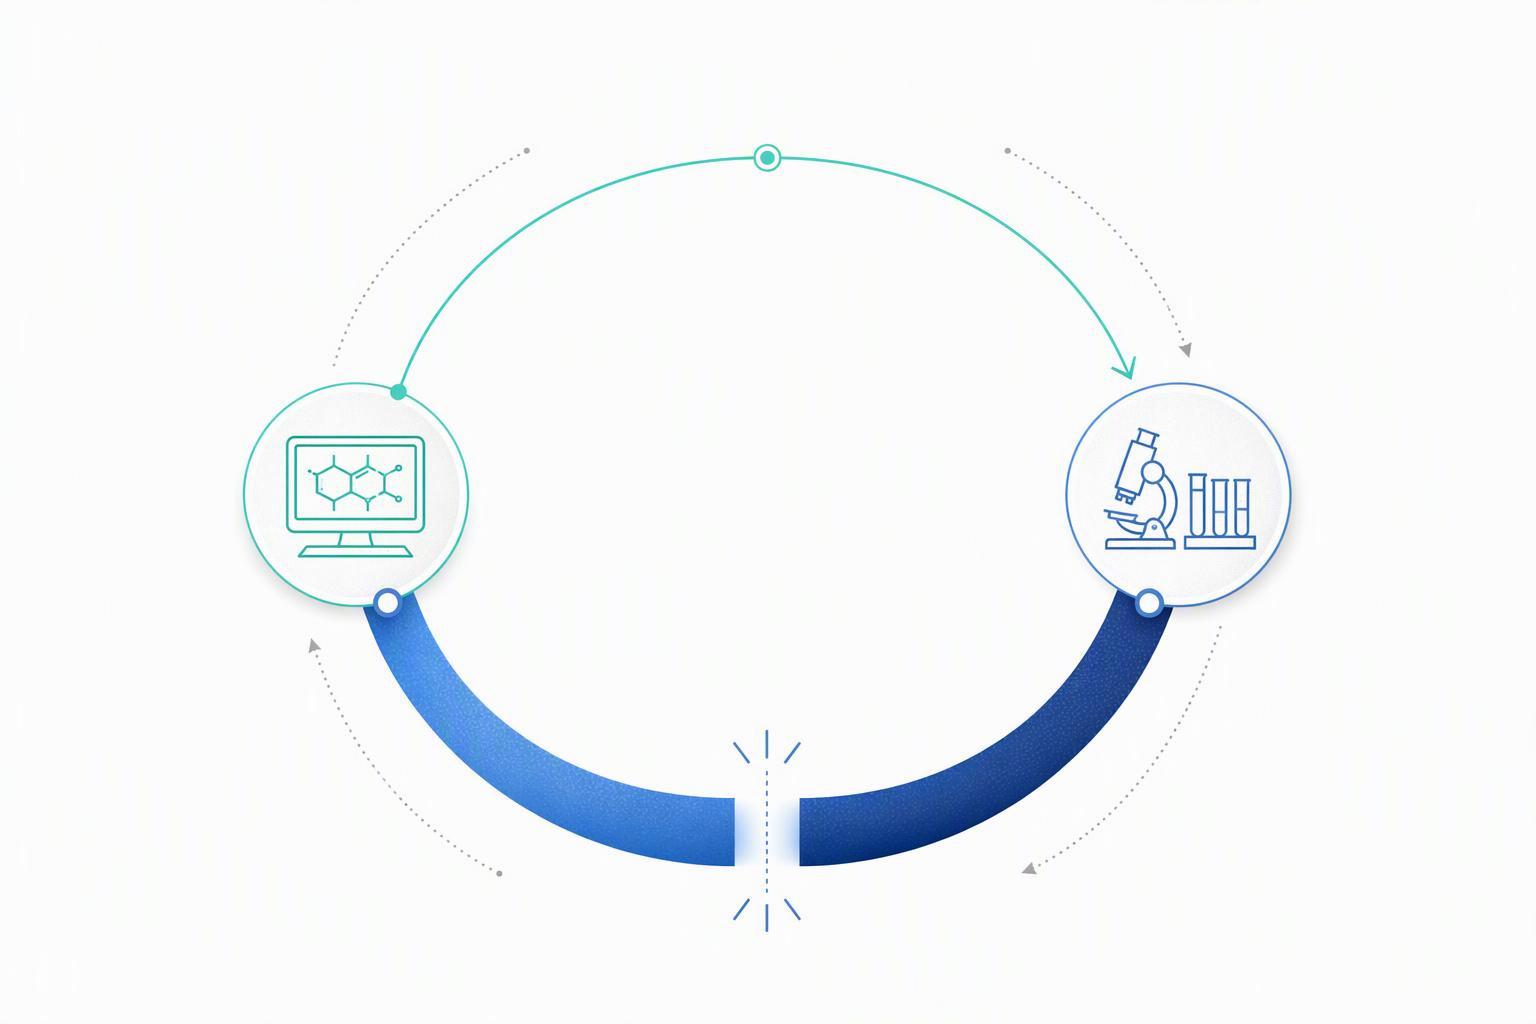
A useful design loop needs credible prediction and credible measurements. Complete records improve interpretation, while model performance, material behavior and biological validity still require independent evaluation.
Audit model validity together with the experimental package. Check independent or leakage-aware evaluation, synthesis success and failure records, identidad, assay controls and the traceability of feedback used for the next design cycle.
The next step is a technical review of one lot’s analytical data package, with the five questions above as the agenda. That is a medium-commitment conversation, and it is the one that determines whether the loop can close.