A narrativa da escassez é o quadro errado para a estratégia do fornecedor de peptídeos
A escassez de peptídeos é real. Tratá-lo como um problema de organização é o que produz más decisões do fornecedor.
Os prazos de entrega relatados para materiais de qualidade farmacêutica passaram de semanas para meses, com PeptideStaff relatando um 60% aumento médio nos prazos de entrega dos aminoácidos Fmoc entre o quarto trimestre 2024 e Q2 2026 e derivativos especiais cotados em 24 para 36 semanas de fornecedores de fonte única. Esse número é informado pelo fornecedor e chega sem uma metodologia publicada, o que importa: quase todos os números quantitativos da cadeia de fornecimento que circulam sobre este tópico remontam a uma família de editores e suas recapitulações posteriores. A narrativa está sendo amplificada mais rápido do que evidenciada.
A escassez é um sintoma. A condição subjacente é que a maioria dos programas de peptídeos nunca projetou sua estratégia de fornecedor em torno do fracasso, então cada interrupção desencadeia uma confusão em vez de uma decisão. A resiliência da cadeia de fornecimento de peptídeos é o que você constrói com antecedência, não o que você monta durante uma ruptura de estoque.
Principal vantagem: Construa sua estratégia de fornecedor de peptídeos com base em cinco critérios de decisão: fornecimento secundário qualificado, prazos de entrega transparentes, controles de processo documentados, caminhos de fabricação escaláveis, e planejamento analítico inicial. Preço e simplicidade de fonte única não são critérios; são padrões que falham sob estresse.
A Visão Convencional: Comprar por preço, Qualifique-se tarde, Fonte única para simplicidade
Esse enquadramento estabelece a posição contra a qual argumenta, que trata a seleção de fornecedores como um exercício comercial com um apêndice técnico. Você executa uma solicitação de proposta, comparar custos unitários e prazos, escolha o fornecedor com melhor pontuação, e, em seguida, entregue o arquivo ao controle de qualidade para fechar a papelada de qualificação. Sob este modelo, a fonte secundária qualificada é uma contingência que você ativa se algo der errado, não é um requisito de design que você financia desde o início.
Neuland Labs afirma a divisão convencional mais explicitamente em seu estrutura para gerenciar dual sourcing com CDMOs, publicado em 27 Fevereiro 2026. A estrutura atribui aproximadamente 70 para 90 por cento do fornecimento para um CDMO primário e 10 para 30 por cento para um secundário, mantém especificações idênticas de qualidade e processo, e geralmente adia a qualificação de desempenho do processo secundário até que o primário seja validado. Também coloca a questão do custo-benefício da dupla fonte no final da Fase II, Fase III, ou comercial, e admite que manter um segundo fornecedor aumenta o tempo de entrega e as despesas gerais de gestão e pode prejudicar o retorno do investimento em volumes baixos.
Essa concessão é o cerne honesto do caso convencional, e explica por que a fonte única persiste. Um fornecedor significa um conjunto de especificações, uma conversa de controle de mudanças, e um relacionamento para gerenciar. A sobrecarga é real, e em pequenos volumes a aritmética muitas vezes favorece a simplicidade.
Por que a visão convencional falha: Três problemas estruturais
The conventional view treats lead times, documentation and capacity as commercial problems to be negotiated. Each is a design problem that procurement cannot solve after the fact.
Lead times and prices have moved in ways that punish late planning. PeptideStaff's 2026 supply-chain review reports that Fmoc amino acid lead times rose an average of 60% between Q4 2024 e Q2 2026, and that specialty derivatives now run 24 para 36 semanas de fornecedores de fonte única. The same review found that resin lead times moved from weeks to months, with pharmaceutical-grade Wang and Rink amide resin extending from 4 para 6 weeks in 2024 para 14 para 20 weeks in 2026 for large lots. Coupling-reagent prices rose 25 para 50 percent over the same window, attributed to reagent production-capacity constraints. These are vendor-published estimates without a stated methodology, so treat the direction as the signal and the exact percentages as indicative.
Documentation gaps, not price, are what actually stall qualification. Bachem’s analysis of amino-acid-derivative sourcing identifies amino-acid-derivative documentation gaps are a known qualification bottleneck: inadequate certificates of analysis, missing TSE/BSE declarations and fragmented change control “result in prolonged qualification processes,” trigger additional testing, and can delay trial or launch approvals by months. No purchasing team can negotiate its way past a missing TSE/BSE guarantee.
Capacity is the binding constraint, and it is committed years before it exists. Announced peptide CDMO capacity investment crossed $2.4 bilhão in the first five months of 2026 sozinho, yet capacity committed years before it exists remains the norm: large-scale solid-phase peptide synthesis capacity carries 18 para 36 month lead times, and global peptide API utilization sits at roughly 87 para 91% against a sustainable long-run 70 para 75%.
⚠️ Aviso: The utilization and investment aggregates above are vendor-reported and unverified. They indicate pressure on the system, not a precise forecast for any single program.
Leia juntos, the three problems describe one failure mode: a peptide supply chain resilience plan built on negotiation rather than on qualification, documentation and capacity design.
O que os dados realmente mostram: O risco está concentrado, Não distribuído
The headline growth number is not the useful signal. Grand View Research’s peptide and oligonucleotide CDMO market report puts the market at $3.1 bilhão em 2025, $3.5 bilhão em 2026, e $8.1 bilhão por 2033 em um 12.9% CAGR, with peptides taking 66.7% de 2025 revenue and North America 36.3%. A market growing at that rate tells you demand is rising. It tells you nothing about whether your program can secure the inputs it needs.
The structure underneath the growth number is where the risk sits. PeptideStaff’s analysis of raw material supply chain resilience reports that more than 80 percent of protected amino acid supply comes from fewer than ten manufacturers, a concentration figure the source presents without a stated methodology, so treat it as directional rather than audited. Concentration of that kind means a single supplier disruption propagates across every program sourcing from that tier, regardless of how many CDMOs you have contracted.
Lead-time structure compounds it. Adesis’s peptide development timeline benchmarks describe technical transfer at 6 para 18 months per handoff in a fragmented multi-CDMO model, with a formal transfer package adding 3 para 6 meses, analytical method transfer and validation 2 para 4 meses, process qualification 2 para 4 meses, and scale-up troubleshooting another 2 para 6 meses. Total fragmented timelines extend to months 25 para 36 for GMP and Phase I to II production. In a fragmented model, half the transfer time can be documentation.
One figure in circulation does not reconcile with the analyst view. A vendor-published projection cited by PeptideStaff puts the global peptide API market at $47.3 bilhão por 2028, acima de $29.1 bilhão em 2024. That is a different market definition from the CDMO services figure above, and the two are not reconcilable as stated, so this article does not use it. Where sources disagree on scope, the honest move is to name the disagreement rather than average it away.
Leia juntos, the evidence points away from a single approved supplier and toward a portfolio of qualified options with documented transferability. Risk in peptide manufacturing scale-up is concentrated in a handful of upstream manufacturers and in the handoffs between organizations, not distributed evenly across the vendor landscape. A vendor strategy built on the growth number alone optimizes for a market condition. A strategy built on concentration and transfer time optimizes for the failure modes that actually stop programs.
Pilar Um: Fornecimento secundário qualificado como requisito de projeto
A backup supplier is not a purchase order. It is a qualification project with its own timeline, and that timeline has to be designed in before you need it.
Under the Q7A/Q11 supplier evaluation expectation, approval rests on an evaluation that gives adequate evidence the supplier can consistently meet specifications, and the guidance is explicit that “Full analyses should be conducted on at least three batches before reducing in-house testing” (ICH Q7A / FDA). Three batches is not a formality; it is calendar time you cannot compress retroactively.
Volume design follows the same logic. A workable split keeps the primary supplier at roughly 70 para 90 percent and the secondary at 10 para 30 por cento, with identical quality and process specifications on both sides, and cost/benefit justification typically arriving from late Phase II/III or commercial (Neuland Laboratories). Secondary process performance qualification is commonly deferred until after primary validation, which is a scheduling decision, not a quality concession.
O modo de falha é específico: a secondary supplier qualified on paper but never exercised is not a qualified secondary source. Qualified secondary sourcing means the relationship has run real batches, under real specifications, on a known cadence.
Pilar Dois: Prazos de entrega transparentes e o horizonte de planejamento que eles implicam
Plan against the longest credible lead time in the chain, not the average. The gap between those two numbers is where peptide vendor strategy quietly fails.
The published figures are wide and single-sourced, so treat each as a directional signal rather than a precise schedule. One industry report describes resin lead times moving from weeks to months, com um 14 para 20 week range, recuperado 2026-09-10. The same source reports that capacity is committed years before it exists: large-scale solid-phase peptide synthesis at 18 para 36 meses, synthesis equipment at 14 para 22 months order-to-delivery, and industrial lyophilizers at 18 para 24 meses.
The planning consequence is concrete. Adesis advises teams to start the CDMO search 18 para 24 months before the first GMP batch, and buyers are reportedly reserving capacity two to three years ahead. Ask every supplier for the assumptions behind their quoted lead time: which raw material, which line, which quality tier.
Pilar Três: Controles de processos documentados e o custo das lacunas de documentação
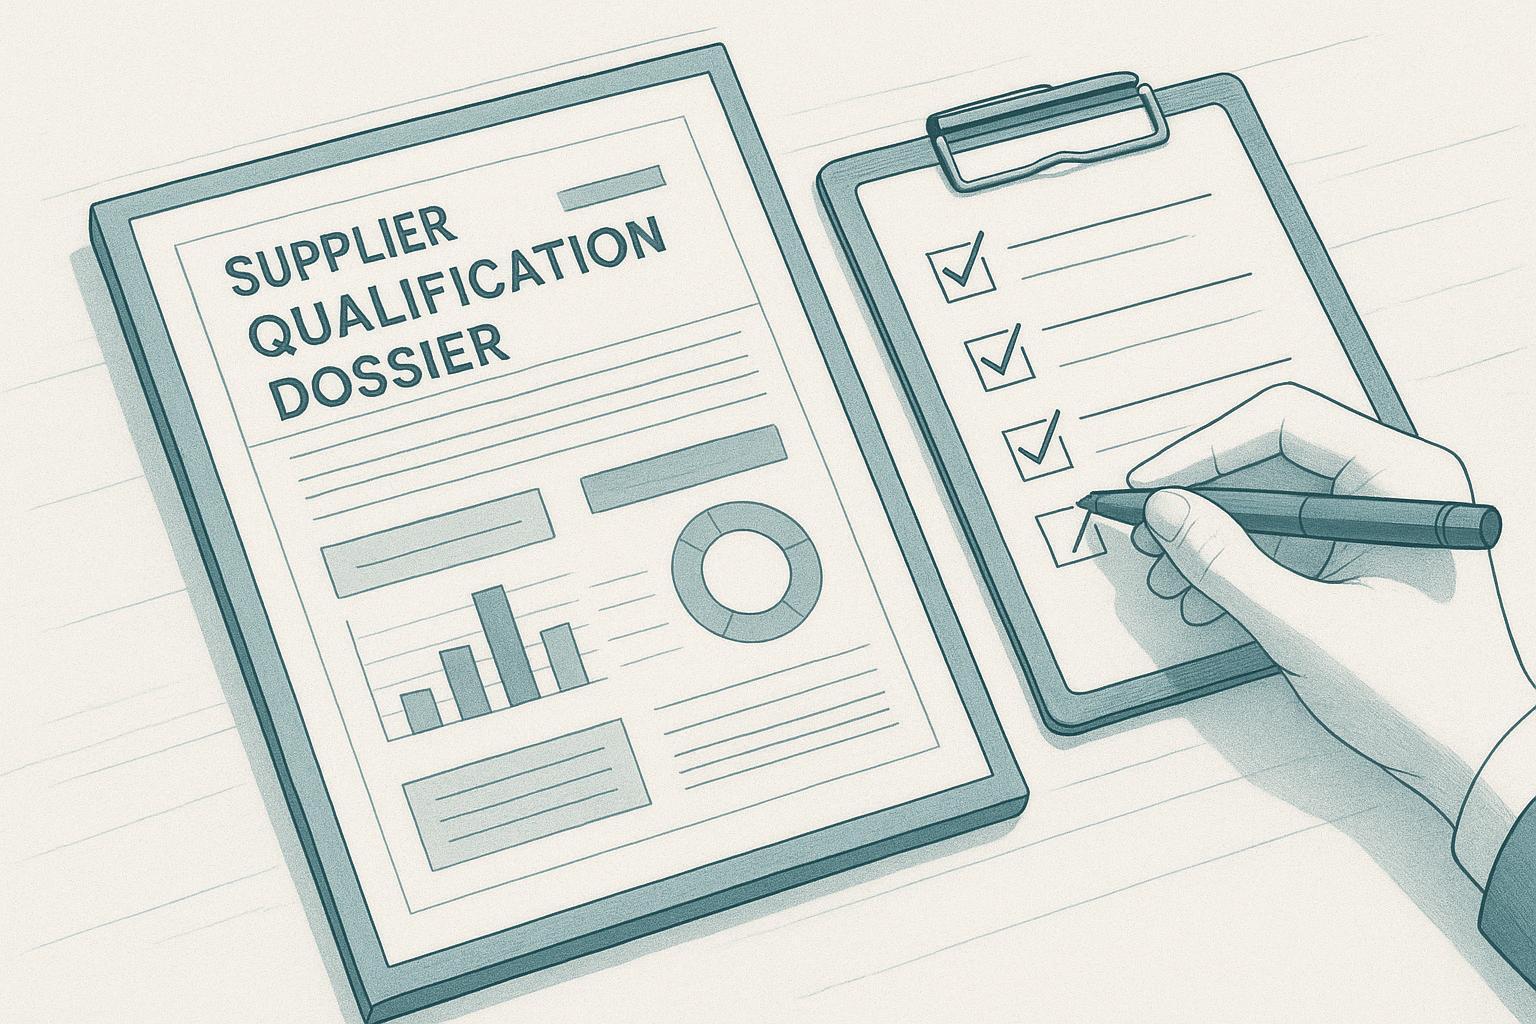
Documentation is a schedule variable, not paperwork. Bachem’s review of GMP amino-acid-derivative sourcing reports that inadequate certificates of analysis, missing TSE/BSE declarations and fragmented change control prolong qualification and can delay approvals by months. The same source notes that 98 percent purity is no longer sufficient, and that validated UHPLC plus orthogonal techniques are now the stated best practice.
The regulatory floor is explicit. Sob ICH Q11, applicants must identify all proposed starting materials, provide specifications, and justify their selection, with Q7 GMP provisions applying from first use of the starting material.
O modo de falha é específico. A tech transfer package that omits counterion exchange conditions, or comparable process detail, does not present as a gap in the package. It surfaces later as a deviation at the receiving site, on the receiving site’s timeline.
Para dica: Review the supplier’s dossier, não apenas o certificado, before selection. Completeness of the CoA, TSE/BSE declarations, change-control records and starting-material justification tells you more about schedule risk than the purity figure does.
Pilar Quatro: Caminhos de fabricação escaláveis decididos antes da expansão
Scale-up risk is a calendar and purification problem, not a chemistry problem. PeptideStaff’s guide to outsourcing peptide scale-up from milligrams to kilograms puts a milligram-to-kilogram project at 12 para 24 meses, and its own phase table sums to 10 para 19 months once CDMO selection, tech transfer, desenvolvimento de processos, pilot batches, the first GMP batch, and release are added up. Those two figures come from the same source and do not agree, which is itself the useful signal: treat any single scale-up timeline as a range to plan against, not a date to commit to. The same source reports that purification accounts for 40 para 60 percent of large-scale cost, and advises starting the CDMO search 18 para 24 months before the first GMP batch is needed.
Síntese de Peptídeos The clinical side compresses differently. PeptideStaff’s analysis of clinical supply manufacturing outsourcing relatórios 6 para 12 months from process transfer to release of clinical supplies, and recommends beginning clinical supply planning 12 para 18 months before the target first-patient-dosed date. It also states that a critical-path failure delays the program by 3 para 6 meses. That last number is the most actionable figure in this section, and it is the one the source does not support with data, so weigh it as an estimate rather than a measured outcome.
The decision this pillar forces is sequencing, not vendor choice. Peptide manufacturing scale-up pathways that get selected after process development has already locked in a synthesis route tend to surface their real cost in purification and analytical method transfer, where the schedule has the least slack. Committing to a pathway early, and confirming that the chosen CDMO can carry it from pilot through GMP, keeps the 18-to-24-month search window from collapsing into a rush qualification.
Pilar Cinco: Planejamento analítico antecipado e transferência de métodos
Analytical work is a sequencing constraint, not a final step. The analytical procedure used for GMP release must already be qualified or validated for its intended use before routine GMP testing is relied on, so method transfer and validation have to be planned before the first GMP lot needs release testing. That requirement is operational, not something the guidelines schedule for you: there is no ICH Q2(R2) rule specifying how many days before a batch to begin. What the guideline does define is the substance of the work. Eu Q2(R2) sets out the validation parameters a peptide purity or impurity method must demonstrate, including specificity and selectivity, precisão, precisão, linearidade, range, LOD, LOQ and robustness, with applicability determined by procedure type and validation planned through a protocol under ICH Q14 that states intended purpose, performance characteristics and acceptance criteria.
Budget for it accordingly. Analytical method transfer and validation typically consume two to four months, which is time that runs in parallel with, não depois, desenvolvimento de processos. A vendor whose synthesis, purification and analytical workflows sit inside one integrated program, as MOL Changes supports, removes one handoff from that sequence, though the same planning discipline applies to any supplier you qualify.
O contra-argumento mais forte: A fonte dupla custa mais do que o risco que mitiga
The objection deserves to be stated without softening: qualifying a second supplier adds lead time, management overhead and a duplicate qualification package, and at low program volume those costs can outweigh the risk they offset. Neuland’s own framework concedes as much, recommending that teams keep the secondary at roughly 10 para 30 percent of volume rather than splitting supply evenly (Neuland Laboratories, recuperado 2026-06-11).
A comparação, no entanto, is not cost versus no cost. It is the cost of a qualified secondary source against the cost of a critical-path failure, and the clinical supply literature puts a single failed campaign can cost three to six months of program delay, with a failed clinical-scale GMP batch running $100K to $500K (clinical supply source, recuperado 2026-06-11).
Concede the boundary honestly: below a certain program volume, or above a certain level of supply certainty, single sourcing is the rational choice, and any framework that prescribes dual sourcing universally is overreaching.
Advertências: Onde este argumento é mais fraco
The strongest objection to this framework is not that it is wrong but that the evidence behind it is thinner than the confidence of its presentation. The widely repeated aggregate figure of $2.4 billion in committed peptide capacity, along with utilization estimates of 87 para 91 por cento, could not be traced to a published methodology, which means the scale of the shortage is asserted rather than measured. The same applies to the claim that more than 80 percent of protected amino acid supply comes from fewer than ten manufacturers: the concentration is plausible and consistent with how the industry describes capacity committed years before it exists, but no source in this review published the underlying supplier data.
That gap matters for how you use the five pillars. They are a planning structure, not a validated predictor. No published dataset shows that programs adopting qualified secondary sourcing, documented process controls or early analytical planning fail less often than programs that do not. What the framework does is make risk visible and assignable: it forces a named owner for each exposure and a decision point before the exposure becomes a crisis. That is a defensible claim even where the outcome data is not, and it is the claim this argument actually rests on.
For some programs, the conventional approach is genuinely correct. If your horizon is short, your material supply has been stable for years, and you have one supplier with a clean audit history and responsive communication, the cost of building a second qualified source may exceed the risk it mitigates. The framework is worth applying where the downside of a supply interruption is measured in months of delay or a lost program, not where it is measured in a rescheduled batch.
The weakest part of this argument is the one I cannot fix with better sourcing: the pillars describe what good risk design looks like, but they do not tell you how much risk reduction each one buys. Treat peptide vendor strategy as a discipline for deciding where to spend attention, not as a formula that produces a number.
Conclusão: Da resposta à escassez ao design de risco
Peptide vendor strategy is a risk-design problem, and the five pillars above are the design: fornecimento secundário qualificado, prazos de entrega transparentes, controles de processo documentados, caminhos de fabricação escaláveis, and analytical readiness decided early.
The shift being called for is a change in when the work happens, not in how much of it there is. Qualificação, documentação, and method planning are cheapest before a program depends on them and most expensive once a timeline does. Peptídeos Sintéticos Treating them as procurement tasks defers that cost into the phase where it does the most damage. Treating them as design requirements moves the same work upstream, where a supplier conversation can still change the outcome.
That reframing is the whole argument. Shortage response reacts to a market that has already moved. Risk design decides, in advance, which failures a program can absorb and which it cannot, then buys accordingly.
If you are structuring a vendor strategy now, talk to an expert about mapping your qualification, lead-time, and documentation requirements against your development timeline. Bring your program stage and your current supplier list; the useful output is a decision structure, not a recommendation.
Disclosure: this article discusses vendor evaluation criteria generally and does not endorse any specific supplier. Produção de Peptídeos
Perguntas frequentes
But doesn’t dual sourcing work only at commercial scale?
Qualification lead time, not volume commitment, é a restrição. The cost-benefit case for a second supplier is usually justified from late Phase II/III or commercial volumes (Neuland Laboratories, recuperado 2026-09-10), and that is where a split such as keeping the secondary at roughly 10 para 30 percent of volume starts to pay for itself. But qualification itself takes months, so the point at which the split makes financial sense and the point at which you must start qualifying are not the same date. Start the qualification work earlier than the volume split justifies.
What if I’ve already committed to a single supplier?
You do not have to start over. The Q7A/Q11 supplier evaluation expectation allows a three-batch, full-analysis approach to build a second supplier’s qualification evidence without duplicating the primary supplier’s work (EU, recuperado 2026-09-10). Budget the transition realistically: transfer packages run 3 para 6 meses, and in a fragmented model, half the transfer time can be documentation (Adesis, recuperado 2026-09-10).
Como você responde às fontes que dizem que a capacidade está se expandindo rápido o suficiente para preencher a lacuna?
Announced capacity is not available capacity. The individual commitments are checkable against company disclosures: Lonza’s CHF 650M for large-scale SPPS at Visp and Geleen, Bachem’s CHF 280M tranche at Sisseln, PolyPeptide’s $180M at Strasbourg, Almac’s £95M, and Thermo Fisher’s $420M across Greenville and Ferentino (PeptídeoStaff, recuperado 2026-09-10). The aggregate figure often quoted alongside them is not verifiable in the same way. And capacity committed years before it exists does not shorten the 18 para 36 month large-scale SPPS lead time that sits between announcement and supply.
O planejamento analítico antecipado realmente muda o cronograma, ou apenas mova o trabalho?
It moves the work off the critical path rather than removing it. The analytical procedure used for GMP release must already be validated for its intended use (EU, recuperado 2026-09-10), which means analytical method transfer and validation consume 2 para 4 months that must precede the first GMP lot’s release testing. No standard specifies a lead time for that sequencing, so the decision is yours to make and defend.