क्या जुलाई 2026 वोट दिया और बदला नहीं
जुलाई में 23-24, 2026, फार्मेसी कंपाउंडिंग सलाहकार समिति ने सात अस्वीकृत पेप्टाइड्स की समीक्षा की और उनमें से छह को 503ए बल्क सूची के लिए अनुशंसित किया। (मैक्डरमोट विल & कस्र्न पत्थर, 2026-07-28). बीपीसी-157, केपीवी, और टीबी-500 प्रत्येक साफ़ हो गया 8-6 एक परहेज के साथ; एमओटीएस-सी उत्तीर्ण 7-5; सेमैक्स पास हो गया 8-5; एपिटलॉन पारित हुआ 7-4; एमिडेलटाइड (डीएसआईपी) असफल 6-7. आरएपीएस एपिटलॉन टैली को इस प्रकार दर्ज किया गया 7-5, इसलिए उस एकल आंकड़े को अस्थिर मानें.
⚠️ चेतावनी: सिफ़ारिश केवल सलाह है. जैसा मिंत्ज़ टिप्पणियाँ, सलाहकार वोट कोई एजेंसी की कार्रवाई नहीं है, और किसी भी लिस्टिंग के लिए अभी भी नोटिस-और-टिप्पणी नियम बनाने की आवश्यकता होती है. कानूनी तौर पर अभी तक कुछ भी नहीं बदला है, और सूची में दिखना एक कंपाउंडिंग-स्थिति प्रश्न है, कोई सुरक्षा खोज नहीं.
जो पहले से लागू है वह अपरिवर्तित है: धारा 503ए के तहत, एक थोक दवा पदार्थ को विश्लेषण के वैध प्रमाण पत्र की आवश्यकता होती है और यह धारा के तहत एफडीए के साथ पंजीकृत प्रतिष्ठान से आना चाहिए 510 (एफडीए, 2026-05-14). एफडीए 503ए सूची को तीन श्रेणियों वाली अंतरिम नीति के तहत निरंतर आधार पर अद्यतन बताता है. यही कारण है कि नीचे दी गई पेप्टाइड दस्तावेज़ीकरण चेकलिस्ट उन रिकॉर्ड्स पर केंद्रित है जिन्हें आप आज नियंत्रित करते हैं.
इस पेप्टाइड दस्तावेज़ीकरण चेकलिस्ट का उपयोग कैसे करें
इस चेकलिस्ट में छह क्षेत्र शामिल हैं: पहचान, पवित्रता, विश्लेषणात्मक तरीकों, अशुद्धता प्रोफाइल, सोर्सिंग, और इच्छित उपयोग. वे कार्य के चार चरणों का मानचित्रण करते हैं: पूर्व-संश्लेषण, प्रक्रिया में, मुक्त करना, और सौंप देना. प्रत्येक आइटम को द्विआधारी निर्णय के रूप में दर्शाया गया है, इसलिए आप हां या ना में जवाब दे सकते हैं और आगे बढ़ सकते हैं.
|
कवरेज क्षेत्र |
पूर्व संश्लेषण |
प्रक्रिया में |
मुक्त करना |
सौंप दो |
|---|---|---|---|---|
|
पहचान |
अनुक्रम और लक्ष्य की पुष्टि की गई |
बिल्डिंग-ब्लॉक रिकॉर्ड |
पहचान की पुष्टि |
अनुक्रम रिकॉर्ड स्थानांतरित किया गया |
|
पवित्रता |
विशिष्टता सेट |
प्रक्रियाधीन जाँच लॉग की गई |
शुद्धता परिणाम बताया गया |
शुद्धता विधि का उल्लेख |
|
विश्लेषणात्मक तरीकों |
विधि चयनित |
विधि लिखित रूप में चलाएँ |
सत्यापन सारांश संलग्न है |
विधि फ़ाइल आपूर्ति की गई |
|
अशुद्धता प्रोफाइल |
ज्ञात अशुद्धियाँ सूचीबद्ध |
प्रोफ़ाइल की निगरानी की गई |
प्रोफ़ाइल रिपोर्ट की गई |
प्रोफ़ाइल डेटा शामिल है |
|
सोर्सिंग |
आपूर्तिकर्ता और लॉट दर्ज किया गया |
सेवाएं हिरासत की श्रृंखला रखी गई |
स्रोत टी दुकान दौड़ने योग्य |
उत्पत्ति प्रलेखित |
|
उपयोग का उद्देश्य |
परिभाषित केस का उपयोग करें |
दायरा अपरिवर्तित |
कथन मिलान का प्रयोग करें |
उपयोग सीमा संसूचित की गई |
दो कार्यक्षेत्र सीमाएँ लागू होती हैं. यह दस्तावेज़ीकरण मार्गदर्शन है, कानूनी या नियामक सलाह नहीं; अनुपालन निर्णय लेने से पहले योग्य नियामक परामर्शदाता से परामर्श लें. और सूची स्थिति और सुरक्षा स्थिति अलग-अलग प्रश्न हैं: किसी सूची में या उसके बाहर दिखाई देने से यह स्वयं स्थापित नहीं होता है कि पेप्टाइड सुरक्षित है या नहीं.
चरणों के माध्यम से क्रम से काम करें, फिर यह पुष्टि करने के लिए कि कुछ भी छूटा नहीं है, अंत में समेकित तालिका का उपयोग करें. निम्नलिखित अनुभाग परिभाषित करते हैं कि प्रत्येक आइटम के लिए पर्याप्त कस्टम पेप्टाइड संश्लेषण दस्तावेज़ीकरण क्या माना जाता है.
चरण 1 - पूर्व-संश्लेषण और सोर्सिंग रिकॉर्ड
जो रिकॉर्ड बनाना सबसे सस्ता होता है उन्हें बाद में दोबारा बनाना सबसे कठिन होता है. इनटेक पर प्राप्त लॉग पर लिखी गई एक लॉट संख्या में कुछ सेकंड लगते हैं; अठारह महीने बाद उसी लॉट संख्या को पुनः प्राप्त करना, जब कोई भागीदार पूछता है कि कौन से बिल्डिंग ब्लॉक एक बैच में गए, इसमें कई दिन लग सकते हैं और ऐसा उत्तर आ सकता है जिसका कोई बचाव नहीं कर सकता.
इस चरण में चार आइटम शामिल हैं, और प्रत्येक एक बाइनरी चेक है:
-
अमीनो एसिड बिल्डिंग ब्लॉक और रेज़िन लॉट नंबर दर्ज किए जाते हैं, प्रत्येक के लिए फ़ाइल पर आपूर्तिकर्ता के सीओए के साथ. ट्रैसेबिलिटी को लॉट स्तर पर जीवित रहना होगा, कैटलॉग स्तर नहीं. आपूर्तिकर्ता योग्यता पैकेट में एक बैच-विशिष्ट सीओए होना चाहिए जो परीक्षणों को रिकॉर्ड करता है, उस लॉट के लिए विशिष्टताएँ और परिणाम, प्लस लॉट और SKU ट्रैसेबिलिटी एक प्रलेखित संश्लेषण मार्ग पर वापस (लियोकेम लैब नोट्स, 2026-06-01). एफडीए की धारा 503ए आवश्यकताओं की अपनी रूपरेखा उसी दिशा की ओर इशारा करती है, वैध सीओए और 510-पंजीकृत निर्माण की उम्मीद है (एफडीए, 2026-05-14).
-
संदर्भ मानक पहचान, लॉट संख्या और समाप्ति का दस्तावेजीकरण किया जाता है.
-
एक लिखित इच्छित-उपयोग विवरण अनुसंधान उपयोग को कंपाउंडिंग से अलग करता है.
-
प्राप्ति से भंडारण तक अभिरक्षा की श्रृंखला प्रलेखित है.
संदर्भ मानक की समाप्ति तिथि आइटम टीमें अक्सर गलत स्थान पर फ़ाइल करती हैं. यह बैच रिकॉर्ड में है, लैब नोटबुक में नहीं, क्योंकि बैच रिकॉर्ड वह है जो एक समीक्षक या भागीदार वास्तव में पढ़ता है. यदि मानक चलने से पहले समाप्त हो गया, इसके विरुद्ध उत्पन्न पहचान डेटा में एक योग्यता होती है जिसे रिकॉर्ड को उसके चेहरे पर बताना चाहिए, और तीन अलमारियों की दूरी पर एक नोटबुक प्रविष्टि सतह पर नहीं आएगी.
चरण 2 - पहचान दस्तावेज़ीकरण
पहचान पवित्रता से अलग प्रश्न है, और ए 98% शुद्धता का आंकड़ा इसका जवाब नहीं देता. शुद्धता परीक्षण यह पुष्टि करता है कि सामग्री का कितना भाग मुख्य शिखर के रूप में उत्सर्जित होता है; यह पुष्टि नहीं करता है कि मुख्य शिखर आपके द्वारा ऑर्डर किया गया पेप्टाइड है. जैसा पेप्ट्रा लैब्स अपने सर्टिफिकेट ऑफ एनालिसिस गाइड में बताती है, शुद्धता परीक्षण पहचान की पुष्टि नहीं है "जब तक कि उस बैच के लिए कोई पहचान परख मौजूद न हो" (2026-05-22). वह सशर्त शब्दांकन ही संपूर्ण मुद्दा है: पहचान को एक ऐसी विधि द्वारा प्रदर्शित किया जाना चाहिए जो आपके अनुक्रम को निकटतम पड़ोसी से अलग कर सके.
तीन विधियाँ वह विशिष्टता रखती हैं. मास स्पेक्ट्रोमेट्री आणविक द्रव्यमान स्थापित करती है, नीचे ±1 Da की इलेक्ट्रोस्प्रे आयनीकरण सहनशीलता के साथ 5,000 इसके ऊपर Da और ±2 Da, और MALDI-TOF छोटे पेप्टाइड्स पर ±0.5 Da धारण करता है (ल्योस लैब्स, 2026-04-08). अमीनो-एसिड विश्लेषण हाइड्रोलिसिस के बाद संरचना की रिपोर्ट करता है, और पेप्टाइड मैपिंग स्वयं अनुक्रम का पता लगाती है. प्रत्येक अपना स्वयं का रिकॉर्ड तैयार करता है: स्पेक्ट्रम या संरचना रिपोर्ट, अपेक्षित बनाम मनाया गया मूल्य, उपकरण और अधिग्रहण पैरामीटर, और विश्लेषक और तारीख.
सत्यापन के दायरे पर, ICH Q2 का वाचन(आर2) वह पेप्टाइड स्टाफ़ पेप्टाइड विधियों पर लागू होता है यह है कि पहचान विधियों को प्रत्येक विशेषता के लिए पूर्ण सत्यापन की आवश्यकता नहीं होती है, लेकिन उन्हें विशिष्टता दिखाने की ज़रूरत है (2026-09-11). इसे विश्राम के बजाय दस्तावेज़ीकरण निर्देश के रूप में पढ़ें: फ़ाइल को अभी भी यह साबित करना है कि विधि भेदभाव करती है, जिसका अर्थ है एक विशिष्ट तर्क, एक प्रतिनिधि स्पेक्ट्रम, और एक घोषित स्वीकृति विंडो. यह वह जगह है जहां पेप्टाइड विश्लेषणात्मक विधि सत्यापन अपेक्षाएं पहचान और परख कार्य के बीच सबसे अधिक भिन्न होती हैं, और जहां एक पतली फ़ाइल का पता लगाना सबसे आसान है.
चरण 3 - शुद्धता दस्तावेज़ीकरण और संख्या का क्या अर्थ है
सीओए शुद्धता का आंकड़ा एक क्षेत्र-प्रतिशत माप है, और यह व्यवस्थित रूप से उच्च पढ़ता है. The record has to state what was measured and how, or the number invites a false conclusion about content.
The distinction is not academic. In one cited metrology example, a manufacturer-stated content of 983.72 mg/g sat against a mass-balance determination of 896.36 ± 0.68 मिलीग्राम/जी, a gap on the order of seventy-plus milligrams per gram (lineara, 2026-05-01). Treat that as a single-upstream illustration, not a general rule.
Water and counter-ions explain much of the difference. Lyophilized peptides are hygroscopic, and failing to correct for water can bias results by several percent; Karl Fischer reports water as % w/w and feeds the net-content and salt-correction calculation (clarascience, 2026-07-15). Counter-ions such as trifluoroacetate, acetate and chloride contribute to batch mass but are not detected at 214 एनएम (peptidetrace, 2026-05-23).
So the record must state the reporting convention: as-is, anhydrous, or salt-free. The 95%, 98% और 99%+ bands are marketing conventions rather than validated specifications; ए 95% specification may contain up to 5% by weight of related substances, synthesis byproducts, residual protecting groups or counter-ions (peptidetrace, 2026-05-23).
कुंजी ले जाएं: An area-percent figure answers “how much of the UV-absorbing material is the main peak”, not “how much peptide is in the vial”.
चरण 4 - विश्लेषणात्मक विधि रिकॉर्ड्स
A method is only reconstructable if the file names the column, the mobile phase, the gradient, the detection wavelength and the integration rules. Everything else is a summary that a receiving lab has to rebuild from scratch.
Conventional peptide practice records a defined parameter set. Column chemistry defaults to C18 for short and mid-length peptides, with C8 for over-retained hydrophobic sequences and 300 Å for longer chains. Dimensions are typically 4.6 × 150 mm for UV purity work and 2.1 × 100 mm for LC-MS, with pore sizes of 100-130 Å and particle sizes of 3.5 या 2.7 माइक्रोन, dropping to 1.7-1.8 µm for LC-MS (reversed-phase HPLC method parameters for peptides, 2026-09-15). Mobile phases run 0.1% टीएफए (मोटे तौर पर 13 मिमी) in water and acetonitrile, switching to 0.1% formic acid for LC-MS. Gradients are scouting at 5-65% B over 60 minutes at 1.0 एमएल/मिनट, or focused at 0.25-0.5% B/min, के साथ 90-95% B wash of at least 5 minutes and re-equilibration of about 10 स्तंभ मात्रा. The measured dwell volume belongs in the method, and detection sits at 214 nm for the peptide bond.
Method status is a separate record from method content. A validated method has been formally demonstrated fit for its intended purpose across the relevant performance characteristics, while a qualified method is a limited demonstration of fitness for a narrower, defined purpose without the full validation evidence (peptide analytical method validation guidelines, 2026-09-11). Which characteristics apply depends on method purpose. Under the adopted मैं Q2(आर2) दिशानिर्देश, असरदार 14 जून 2024, the named characteristics include accuracy, शुद्धता, विशेषता, पता करने की सीमा, मात्रा सीमा, linearity and range. In peptide practice, assay methods carry the full set, with a range of 80-120% of target, recovery of 98-102%, RSD at or below 2% and r² of at least 0.999. Impurity methods lean on specificity and sensitivity instead: a range from the reporting threshold to 120% of the specification limit, a limit of detection well below the reporting threshold, a limit of quantitation at or below it, and forced degradation to establish specificity. Treat those figures as peptide convention rather than as ICH requirements.
|
Method status |
पेप्टाइड संश्लेषण What it demonstrates |
Documentation it requires |
|---|---|---|
|
मान्य |
Fitness for the intended purpose across the relevant performance characteristics |
Full validation protocol, predefined acceptance criteria, कच्चा डेटा, and a validation report covering each assigned characteristic |
|
योग्य |
Limited fitness for a narrower, defined purpose |
Scope statement naming the defined purpose, the characteristics actually tested, परिणाम, and an explicit note of what was not assessed |
System suitability criteria and integration settings are records in their own right, not settings buried in the software. Write down the resolution, पूंछ कारक, retention time repeatability and signal-to-noise thresholds that a run must meet before its result counts, and record सिंथेटिक पेप्टाइड्स the integration rules (baseline mode, smoothing, minimum area threshold, manual intervention and who authorised it). A purity figure produced under unstated integration rules cannot be defended, and a method that cannot be reconstructed cannot be transferred to a larger scale.
चरण 5 - अशुद्धता प्रोफ़ाइल दस्तावेज़ीकरण
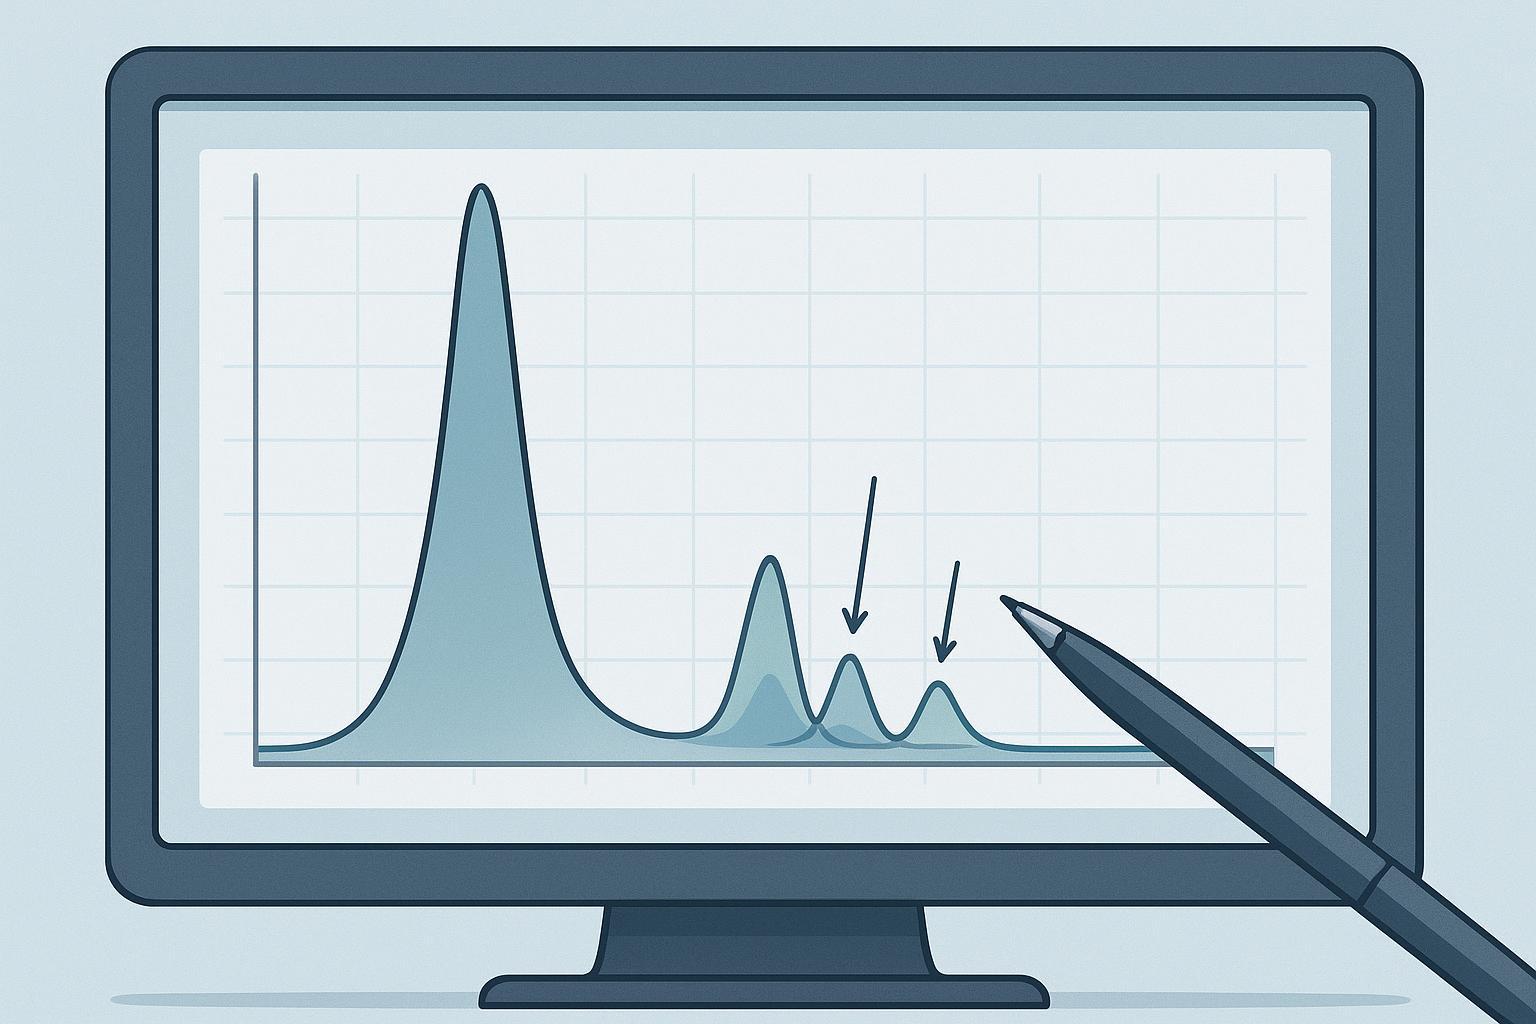
A defensible impurity record lists the classes you looked for and the method that would have found each one. Deletion sequences show as a mass deficit for the absent residue; truncated chains sit at lower mass; oxidised methionine appears as a +16 Da satellite; deamidated Asn or Gln shifts by +1 Da with altered retention; racemisation changes activity without changing mass; TFA adducts show up directly in the chromatogram (lyoslabs, 2026-04-08). Because these species can be close in both mass and retention, a single chromatogram will not separate them all, which is why peptide purity and impurity profile documentation pairs orthogonal methods rather than one run.
Thresholds for reporting, identification and qualification trace to ICH Q3B(आर2), which sets identification typically at 0.10% या 1.0 mg per day intake, whichever is lower. That framework does not apply to research-grade material in the same formal sense, so state which threshold basis your record uses.
चरण 6 - रिलीज़ और हैंडओवर रिकॉर्ड्स
Assemble the hand-over as a package, not as a folder of files. The package is the artifact a partner actually reads, and a missing element blocks a program even when the underlying work was sound. Start with the peptide certificate of analysis requirements: the product name must match the vial label, the batch or lot must match both the vial and the order paperwork, and the report must carry the test date, the analytical method, the batch-specific purity result, the testing laboratory, and a lab-controlled public report URL rather than a supplier-hosted page alone (Peptra Labs, 2026-05-22). पेप्टाइड उत्पादन
Bind every reported value to the raw data behind it, and keep the version and approval trail with the record set. Records should be attributable, पढ़ने योग्य, समसामयिक, मौलिक और सटीक, and the set should let a reviewer reconstruct the batch without contacting the lab.
टिप के लिए: If a reviewer has to email you to understand a number, the package is incomplete. के बारे में
Documentation packages of this kind are the workflow एमओएल परिवर्तन का समर्थन करता है.
समेकित पेप्टाइड दस्तावेज़ीकरण चेकलिस्ट
This peptide documentation checklist condenses the six coverage areas into one printable page. Each row is a binary judgment: either the record exists in the lot file or it does not. Hand it to the team before the next campaign starts, and treat any “no” as an open item rather than a formality.
|
चरण |
चेकलिस्ट आइटम |
Record that satisfies it |
हाँ / नहीं |
|---|---|---|---|
|
1. Pre-synthesis and sourcing |
Is the resin, each amino acid, and each reagent lot identified with supplier and lot number? |
Sourcing log with supplier certificates |
|
|
1. Pre-synthesis and sourcing |
Is the intended sequence, including any non-natural or modified residues, written down before synthesis begins? |
Approved target sequence sheet |
|
|
1. Pre-synthesis and sourcing |
Is the counterion and salt form specified up front? |
Specification sheet |
|
|
2. पहचान |
Is the peptide confirmed by mass spectrometry against the theoretical mass? |
Mass spectrum with calculated versus observed mass |
|
|
2. पहचान |
Is the primary sequence confirmed independently of mass alone? |
Sequencing or MS/MS fragmentation data |
|
|
2. पहचान |
Are the chromatographic identity markers recorded? |
Analytical HPLC retention data |
|
|
3. पवित्रता |
Is the purity figure stated with its method, स्तंभ, और तरंग दैर्ध्य का पता लगाना? |
HPLC chromatogram and method reference |
|
|
3. पवित्रता |
Is the purity expressed on a defined basis, with counterion and water content stated separately? |
Purity calculation sheet |
|
|
3. पवित्रता |
Is the integration method and any peak exclusion rule documented? |
Integration report |
|
|
4. Analytical method records |
Is the method described well enough for another laboratory to reproduce it? |
Method file with column, मोबाइल फेज़, ग्रेडियेंट, प्रवाह, and detection |
|
|
4. Analytical method records |
Is method validation or verification data on file for the intended use? |
Validation or verification report |
|
|
4. Analytical method records |
Are system suitability criteria defined and met on the day of analysis? |
System suitability record |
|
|
5. अशुद्धता प्रोफ़ाइल |
Are related substances reported individually rather than as one lumped figure? |
Impurity table with retention times and relative amounts |
|
|
5. अशुद्धता प्रोफ़ाइल |
Are specified, अनिर्दिष्ट, and total impurity limits stated? |
Specification with impurity thresholds |
|
|
5. अशुद्धता प्रोफ़ाइल |
Are residual solvents, प्रतिवाद सामग्री, and water addressed? |
Residual solvent and content data |
|
|
6. Release and hand-over |
Does the certificate of analysis match the batch record values exactly? |
CoA cross-checked against raw data |
|
|
6. Release and hand-over |
Is the stability or retest date supported by data rather than assumed? |
Stability summary or retest justification |
|
|
6. Release and hand-over |
Is the full data package transferable to a partner or regulator without further explanation? |
Complete lot file index |
Two habits keep this page useful. पहला, review it at the point each record is generated, not at release, because reconstructing an integration rule or a system suitability result months later is where most gaps appear. दूसरा, keep the raw data behind every row, since a summary table without the underlying chromatogram or spectrum does not survive scrutiny.
कुंजी ले जाएं: A documentation package is only as strong as its weakest row. If any item above cannot be answered with a specific record, that is the item to fix before the next lot is released.
पेप्टाइड दस्तावेज़ीकरण के बारे में सामान्य प्रश्न
पेप्टाइड दस्तावेज़ीकरण को कितने समय तक रखा जाना चाहिए?
Retain records for as long as the material can be in use or referenced, then add a defined margin on top of that. Because a lot may be held, re-tested, or shipped years after release, the retention period should be set by the longest plausible use window, not by the release date. Where a lot is tied to a sterility or endotoxin claim, the retention clock follows the longest applicable requirement, since those records are the ones most likely to be requested later.
What if a supplier’s certificate of analysis omits a method detail?
Ask the supplier for the missing detail in writing before you accept the lot, and record the request and the response. A certificate that reports a purity figure without naming the method, स्तंभ, or detection wavelength cannot be independently interpreted, so the gap is a documentation defect rather than a formatting preference. If the supplier will not supply it, treat the lot as unverified for that attribute and say so in your own records.
क्या किसी दिए गए उद्देश्य के लिए एक योग्य विधि स्वीकार्य है?
हाँ, provided the qualification is documented against the intended use and the limits are stated. A qualified method is acceptable when its fitness for the specific purpose has been demonstrated and recorded; it is not acceptable as a substitute for a validated method where the purpose requires validated performance. The deciding question is what the result will be used to support, not how the method was labeled.
जब बहुत सारी सिस्टम उपयुक्तता विफल हो जाती है तो हमें क्या करना चाहिए??
Do not release the lot, and do not re-integrate or re-run until the failure is investigated and the cause is identified. A system suitability failure is a signal about the method or the instrument as much as about the sample, so the investigation should establish which before any repeat injection is treated as a valid result. Document the failure, the investigation, and the resolution together, because the failed run is part of the lot’s record.
कुंजी ले जाएं: Each answer above stands alone, so you can lift the one you need into a supplier query, an SOP note, or a review meeting without losing its context.
अगले कदम
The regulatory outcome is not yours to control; the record set is. Start tomorrow with the phase that is weakest in your current files, usually Phase 4, and work forward. A peptide documentation checklist only helps if someone can hand the file to a reviewer who was not in the room.
If you would like a worked example, MOL Changes supports teams with a sample documentation package covering method files, impurity tables and lot-level release records, which can be used to benchmark your own template before your next audit cycle.
This article is documentation guidance, कानूनी या नियामक सलाह नहीं. Consult qualified regulatory counsel before making compliance decisions. Reviewed by the MOL Changes technical documentation team. MOL Changes provides custom peptide synthesis and analytical services; this article is educational and does not constitute a product recommendation.