Che luglio 2026 Il voto è cambiato e non è cambiato
A luglio 23-24, 2026, il comitato consultivo sui composti farmaceutici ha esaminato sette peptidi non approvati e ne ha raccomandati sei per l'elenco dei prodotti sfusi 503A (McDermott Will & Smeriglio, 2026-07-28). BPC-157, KPV, e TB-500 eliminati ciascuno 8-6 con una astensione; MOT-c superato 7-5; Semax è passato 8-5; Epitalon è passato 7-4; Emideltide (DSIP) fallito 6-7. RAP ha registrato il conteggio dell'Epitalon come 7-5, quindi tratta quella singola figura come instabile.
⚠️ Attenzione: La raccomandazione è solo consultiva. COME Mintz note, un voto consultivo non è un'azione dell'agenzia, e qualsiasi elenco richiede ancora una regolamentazione in materia di avvisi e commenti. Dal punto di vista giuridico non è ancora cambiato nulla, e nell'elenco appare una domanda sullo stato della capitalizzazione, non un risultato di sicurezza.
Resta invariato quanto già applicabile: ai sensi della sezione 503A, un farmaco sfuso necessita di un certificato di analisi valido e deve provenire da uno stabilimento registrato presso la FDA nella sezione 510 (FDA, 2026-05-14). La FDA descrive inoltre l'elenco 503A come aggiornato su base continuativa nell'ambito di una politica provvisoria con tre categorie. Questo è il motivo per cui la seguente lista di controllo della documentazione sui peptidi si concentra sui record che controlli oggi.
Come utilizzare questa lista di controllo della documentazione sui peptidi
Questa lista di controllo copre sei aree: identità, purezza, metodi analitici, profili di impurità, approvvigionamento, e destinazione d'uso. Si associano a quattro fasi di lavoro: pre-sintesi, in corso, pubblicazione, e consegna. Ogni elemento è formulato come un giudizio binario, quindi puoi rispondere sì o no e andare avanti.
|
Area di copertura |
Pre-sintesi |
In corso |
Pubblicazione |
Devolvere |
|---|---|---|---|---|
|
Identità |
Sequenza e obiettivo confermati |
Record di elementi costitutivi |
Conferma dell'identità |
Record di sequenza trasferito |
|
Purezza |
Insieme delle specifiche |
Controlli in-process registrati |
Risultato di purezza dichiarato |
Metodo di purezza citato |
|
Metodi analitici |
Metodo selezionato |
Metodo eseguito come scritto |
Riepilogo della convalida allegato |
File del metodo fornito |
|
Profili di impurità |
Impurezze note elencate |
Profilo monitorato |
Profilo segnalato |
Dati del profilo inclusi |
|
Approvvigionamento |
Fornitore e lotto registrati |
Servizi Catena di custodia mantenuta |
Fonte t Negozio corsabile |
Provenienza documentata |
|
Destinazione d'uso |
Caso d'uso definito |
Ambito invariato |
Utilizza corrispondenze di istruzioni |
Limiti d'uso comunicati |
Si applicano due limiti di ambito. Questa è una guida alla documentazione, non consulenza legale o normativa; consultare un consulente normativo qualificato prima di prendere decisioni in materia di conformità. E lo stato dell'elenco e lo stato di sicurezza sono domande separate: la comparsa o meno di un elenco non stabilisce di per sé se un peptide è sicuro.
Lavora attraverso le fasi in ordine, quindi utilizzare la tabella consolidata alla fine per confermare che non è stato tralasciato nulla. Le sezioni che seguono definiscono ciò che conta come documentazione adeguata sulla sintesi peptidica personalizzata per ciascun articolo.
Fase 1 — Documenti di pre-sintesi e di approvvigionamento
I documenti più economici da creare sono i più difficili da ricostruire in seguito. La scrittura del numero di lotto su un registro di ricezione al momento dell'immissione richiede pochi secondi; recuperando lo stesso numero di lotto diciotto mesi dopo, quando un partner chiede quali elementi costitutivi sono stati inseriti in un batch, possono volerci giorni e finire con una risposta che nessuno può difendere.
Quattro elementi appartengono a questa fase, e ciascuno è un controllo binario:
-
Vengono registrati i numeri di lotto dei componenti di amminoacidi e delle resine, con il CoA del fornitore in archivio per ciascuno. La tracciabilità deve sopravvivere a livello di lotto, non a livello di catalogo. Un pacchetto di qualificazione del fornitore dovrebbe contenere un certificato di autenticità specifico del lotto che registri i test, specifiche e risultati per quel lotto, inoltre la tracciabilità di lotti e SKU risale a un percorso di sintesi documentato (appunti del laboratorio Lyochem, 2026-06-01). Lo schema dei requisiti della sezione 503A della FDA punta nella stessa direzione, aspettandosi un CoA valido e una produzione registrata 510 (FDA, 2026-05-14).
-
Identità standard di riferimento, numero di lotto e scadenza sono documentati.
-
Una dichiarazione scritta sulla destinazione d'uso distingue l'uso per la ricerca dal compounding.
-
La catena di custodia dal ricevimento allo stoccaggio è documentata.
La data di scadenza dello standard di riferimento è l’elemento che i team più spesso archiviano nel posto sbagliato. Appartiene al record batch, non in un quaderno di laboratorio, perché il batch record è ciò che effettivamente legge un revisore o un partner. Se lo standard è scaduto prima della corsa, i dati di identità generati contro di esso comportano una qualifica che il record dovrebbe riportare sul suo volto, e una voce di taccuino a tre scaffali di distanza non lo farà emergere.
Fase 2 — Documentazione d'identità
L’identità è una questione separata dalla purezza, e un 98% la cifra della purezza non risponde. I test di purezza confermano la quantità di materiale che eluisce come picco principale; non conferma che il picco principale sia il peptide ordinato. COME Peptra Labs lo spiega nella sua guida al certificato di analisi, il test di purezza non è una conferma dell’identità “a meno che non esista un test di identità per quel lotto” (2026-05-22). Quella formulazione condizionale è il punto centrale: l'identità deve essere dimostrata con un metodo in grado di distinguere la tua sequenza da quella più vicina.
Tre metodi portano questa specificità. La spettrometria di massa stabilisce la massa molecolare, con tolleranze di ionizzazione elettrospray di ±1 Da di seguito 5,000 Da e ±2 Da sopra, e MALDI-TOF che tiene ± 0,5 Da su peptidi più corti (Laboratori Lyos, 2026-04-08). L'analisi degli amminoacidi riporta la composizione dopo l'idrolisi, and peptide mapping locates the sequence itself. Each produces its own record: the spectrum or composition report, the expected versus observed value, the instrument and acquisition parameters, and the analyst and date.
On validation scope, the reading of ICH Q2(R2) that Peptide Staff applies to peptide methods is that identity methods do not need full validation across every characteristic, but they do need to show specificity (2026-09-11). Read that as a documentation instruction rather than a relaxation: the file still has to prove the method discriminates, which means a specificity rationale, a representative spectrum, and a stated acceptance window. This is where peptide analytical method validation expectations differ most sharply between identity and assay work, and where a thin file is easiest to spot.
Fase 3 - Documentazione sulla purezza e significato del numero
A CoA purity figure is an area-percent measure, and it systematically reads high. The record has to state what was measured and how, or the number invites a false conclusion about content.
The distinction is not academic. In one cited metrology example, a manufacturer-stated content of 983.72 mg/g sat against a mass-balance determination of 896.36 ± 0.68 mg/g, a gap on the order of seventy-plus milligrams per gram (lineara, 2026-05-01). Treat that as a single-upstream illustration, not a general rule.
Water and counter-ions explain much of the difference. Lyophilized peptides are hygroscopic, and failing to correct for water can bias results by several percent; Karl Fischer reports water as % w/w and feeds the net-content and salt-correction calculation (clarascience, 2026-07-15). Counter-ions such as trifluoroacetate, acetate and chloride contribute to batch mass but are not detected at 214 nm (peptidetrace, 2026-05-23).
So the record must state the reporting convention: as-is, anhydrous, or salt-free. IL 95%, 98% E 99%+ bands are marketing conventions rather than validated specifications; UN 95% specification may contain up to 5% by weight of related substances, synthesis byproducts, residual protecting groups or counter-ions (peptidetrace, 2026-05-23).
Chiave da asporto: An area-percent figure answers “how much of the UV-absorbing material is the main peak”, not “how much peptide is in the vial”.
Fase 4 — Documenti sui metodi analitici
A method is only reconstructable if the file names the column, the mobile phase, the gradient, the detection wavelength and the integration rules. Everything else is a summary that a receiving lab has to rebuild from scratch.
Conventional peptide practice records a defined parameter set. Column chemistry defaults to C18 for short and mid-length peptides, with C8 for over-retained hydrophobic sequences and 300 Å for longer chains. Dimensions are typically 4.6 × 150 mm for UV purity work and 2.1 × 100 mm for LC-MS, with pore sizes of 100-130 Å and particle sizes of 3.5 O 2.7 µm, dropping to 1.7-1.8 µm for LC-MS (reversed-phase HPLC method parameters for peptides, 2026-09-15). Mobile phases run 0.1% TFA (all'incirca 13 mm) in water and acetonitrile, switching to 0.1% formic acid for LC-MS. Gradients are scouting at 5-65% B over 60 minuti a 1.0 ml/min, or focused at 0.25-0.5% B/min, con a 90-95% B wash of at least 5 minutes and re-equilibration of about 10 column volumes. The measured dwell volume belongs in the method, and detection sits at 214 nm for the peptide bond.
Method status is a separate record from method content. A validated method has been formally demonstrated fit for its intended purpose across the relevant performance characteristics, while a qualified method is a limited demonstration of fitness for a narrower, defined purpose without the full validation evidence (peptide analytical method validation guidelines, 2026-09-11). Which characteristics apply depends on method purpose. Under the adopted Io Q2(R2) linea guida, efficace 14 Giugno 2024, the named characteristics include accuracy, precisione, specificità, limite di rilevamento, limite di quantificazione, linearità e portata. In peptide practice, assay methods carry the full set, with a range of 80-120% of target, recovery of 98-102%, RSD at or below 2% and r² of at least 0.999. Impurity methods lean on specificity and sensitivity instead: a range from the reporting threshold to 120% del limite di specifica, a limit of detection well below the reporting threshold, a limit of quantitation at or below it, and forced degradation to establish specificity. Treat those figures as peptide convention rather than as ICH requirements.
|
Method status |
Sintesi peptidica What it demonstrates |
Documentation it requires |
|---|---|---|
|
Validated |
Fitness for the intended purpose across the relevant performance characteristics |
Full validation protocol, predefined acceptance criteria, dati grezzi, and a validation report covering each assigned characteristic |
|
Qualificato |
Limited fitness for a narrower, defined purpose |
Scope statement naming the defined purpose, the characteristics actually tested, risultati, and an explicit note of what was not assessed |
System suitability criteria and integration settings are records in their own right, not settings buried in the software. Write down the resolution, fattore di scodamento, retention time repeatability and signal-to-noise thresholds that a run must meet before its result counts, and record Peptidi sintetici the integration rules (baseline mode, smoothing, minimum area threshold, manual intervention and who authorised it). A purity figure produced under unstated integration rules cannot be defended, and a method that cannot be reconstructed cannot be transferred to a larger scale.
Fase 5 — Documentazione sul profilo delle impurità
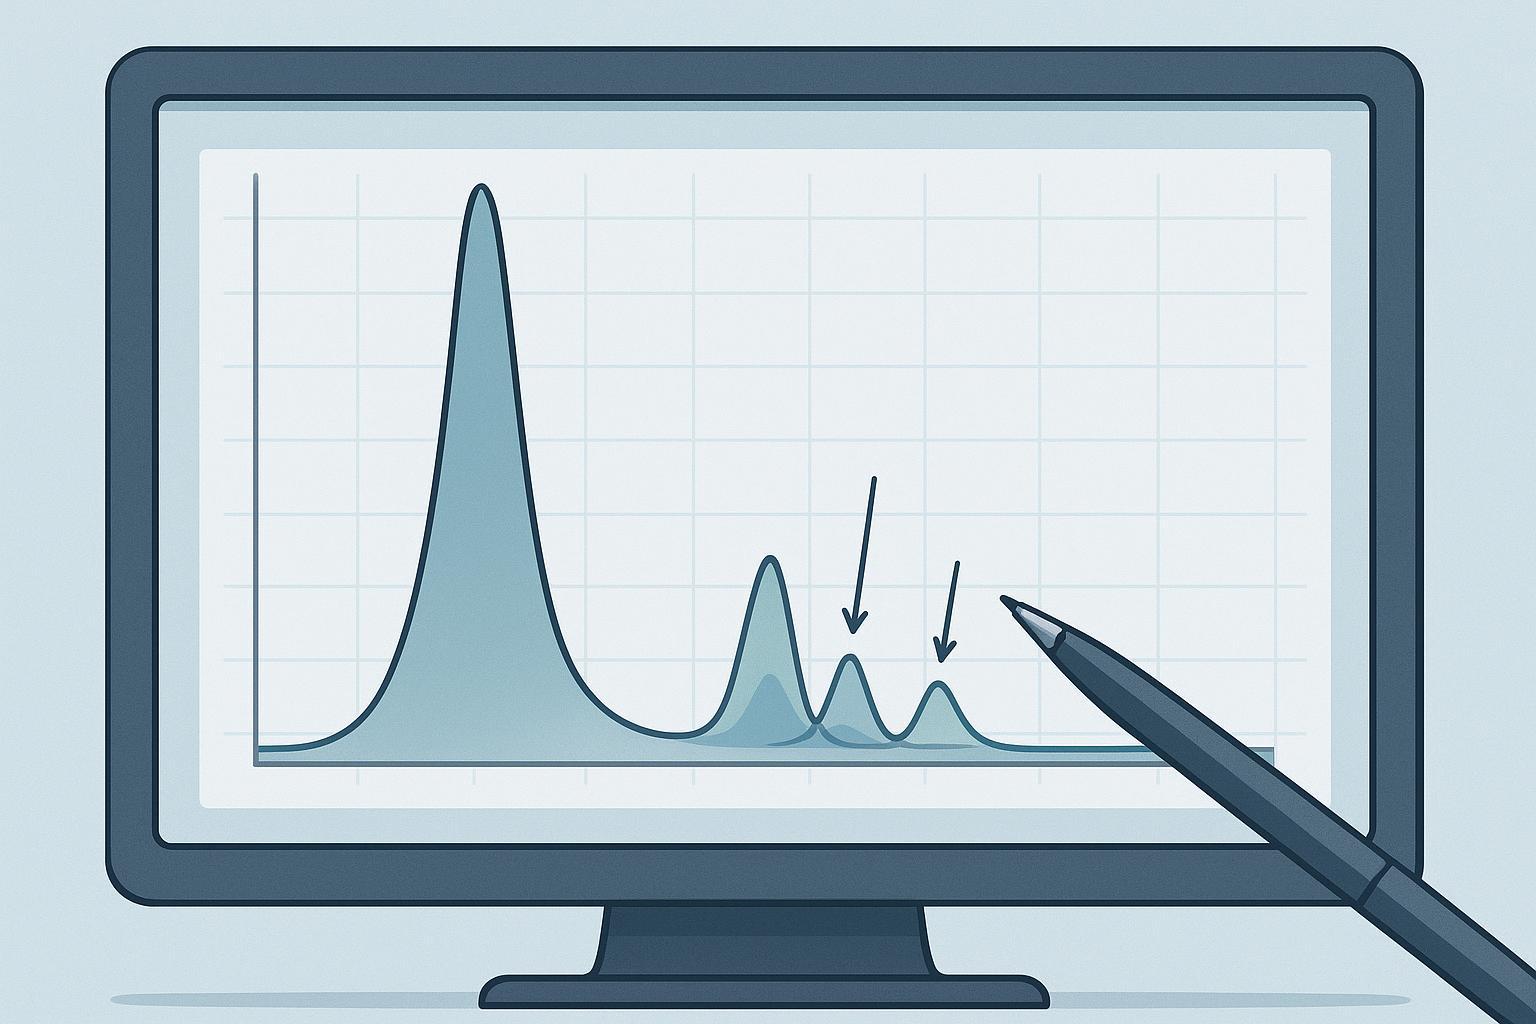
A defensible impurity record lists the classes you looked for and the method that would have found each one. Deletion sequences show as a mass deficit for the absent residue; truncated chains sit at lower mass; oxidised methionine appears as a +16 Da satellite; deamidated Asn or Gln shifts by +1 Da with altered retention; racemisation changes activity without changing mass; TFA adducts show up directly in the chromatogram (lyoslabs, 2026-04-08). Because these species can be close in both mass and retention, a single chromatogram will not separate them all, which is why peptide purity and impurity profile documentation pairs orthogonal methods rather than one run.
Thresholds for reporting, identification and qualification trace to ICH Q3B(R2), which sets identification typically at 0.10% O 1.0 mg per day intake, whichever is lower. That framework does not apply to research-grade material in the same formal sense, so state which threshold basis your record uses.
Fase 6 — Documenti di rilascio e consegna
Assemble the hand-over as a package, not as a folder of files. The package is the artifact a partner actually reads, and a missing element blocks a program even when the underlying work was sound. Start with the peptide certificate of analysis requirements: the product name must match the vial label, the batch or lot must match both the vial and the order paperwork, and the report must carry the test date, the analytical method, the batch-specific purity result, il laboratorio di prova, and a lab-controlled public report URL rather than a supplier-hosted page alone (Peptra Labs, 2026-05-22). Produzione di peptidi
Bind every reported value to the raw data behind it, and keep the version and approval trail with the record set. Records should be attributable, leggibile, contemporaneo, originale e accurato, and the set should let a reviewer reconstruct the batch without contacting the lab.
Per Suggerimento: If a reviewer has to email you to understand a number, the package is incomplete. Di
Documentation packages of this kind are the workflow Modifiche MOL supporta.
Lista di controllo consolidata della documentazione sui peptidi
This peptide documentation checklist condenses the six coverage areas into one printable page. Each row is a binary judgment: either the record exists in the lot file or it does not. Hand it to the team before the next campaign starts, and treat any “no” as an open item rather than a formality.
|
Fase |
Voce della lista di controllo |
Record that satisfies it |
SÌ / NO |
|---|---|---|---|
|
1. Pre-synthesis and sourcing |
Is the resin, each amino acid, and each reagent lot identified with supplier and lot number? |
Sourcing log with supplier certificates |
|
|
1. Pre-synthesis and sourcing |
Is the intended sequence, including any non-natural or modified residues, written down before synthesis begins? |
Approved target sequence sheet |
|
|
1. Pre-synthesis and sourcing |
Is the counterion and salt form specified up front? |
Specification sheet |
|
|
2. Identità |
Is the peptide confirmed by mass spectrometry against the theoretical mass? |
Mass spectrum with calculated versus observed mass |
|
|
2. Identità |
Is the primary sequence confirmed independently of mass alone? |
Sequencing or MS/MS fragmentation data |
|
|
2. Identità |
Are the chromatographic identity markers recorded? |
Analytical HPLC retention data |
|
|
3. Purezza |
Is the purity figure stated with its method, colonna, e lunghezza d'onda di rilevamento? |
HPLC chromatogram and method reference |
|
|
3. Purezza |
Is the purity expressed on a defined basis, with counterion and water content stated separately? |
Purity calculation sheet |
|
|
3. Purezza |
Is the integration method and any peak exclusion rule documented? |
Integration report |
|
|
4. Analytical method records |
Is the method described well enough for another laboratory to reproduce it? |
Method file with column, fase mobile, pendenza, fluire, and detection |
|
|
4. Analytical method records |
Is method validation or verification data on file for the intended use? |
Validation or verification report |
|
|
4. Analytical method records |
Are system suitability criteria defined and met on the day of analysis? |
System suitability record |
|
|
5. Profilo di impurità |
Are related substances reported individually rather than as one lumped figure? |
Impurity table with retention times and relative amounts |
|
|
5. Profilo di impurità |
Are specified, non specificato, and total impurity limits stated? |
Specification with impurity thresholds |
|
|
5. Profilo di impurità |
Are residual solvents, contenuto di controione, and water addressed? |
Residual solvent and content data |
|
|
6. Release and hand-over |
Does the certificate of analysis match the batch record values exactly? |
CoA cross-checked against raw data |
|
|
6. Release and hand-over |
Is the stability or retest date supported by data rather than assumed? |
Stability summary or retest justification |
|
|
6. Release and hand-over |
Is the full data package transferable to a partner or regulator without further explanation? |
Complete lot file index |
Two habits keep this page useful. Primo, review it at the point each record is generated, not at release, because reconstructing an integration rule or a system suitability result months later is where most gaps appear. Secondo, keep the raw data behind every row, since a summary table without the underlying chromatogram or spectrum does not survive scrutiny.
Chiave da asporto: A documentation package is only as strong as its weakest row. If any item above cannot be answered with a specific record, that is the item to fix before the next lot is released.
Domande comuni sulla documentazione dei peptidi
Per quanto tempo deve essere conservata la documentazione relativa ai peptidi?
Retain records for as long as the material can be in use or referenced, then add a defined margin on top of that. Because a lot may be held, re-tested, or shipped years after release, the retention period should be set by the longest plausible use window, not by the release date. Where a lot is tied to a sterility or endotoxin claim, the retention clock follows the longest applicable requirement, since those records are the ones most likely to be requested later.
What if a supplier’s certificate of analysis omits a method detail?
Ask the supplier for the missing detail in writing before you accept the lot, and record the request and the response. A certificate that reports a purity figure without naming the method, colonna, or detection wavelength cannot be independently interpreted, so the gap is a documentation defect rather than a formatting preference. If the supplier will not supply it, treat the lot as unverified for that attribute and say so in your own records.
Un metodo qualificato è accettabile per un determinato scopo?
SÌ, provided the qualification is documented against the intended use and the limits are stated. A qualified method is acceptable when its fitness for the specific purpose has been demonstrated and recorded; it is not acceptable as a substitute for a validated method where the purpose requires validated performance. The deciding question is what the result will be used to support, not how the method was labeled.
Cosa dovremmo fare quando molte cose non sono adeguate all’idoneità del sistema?
Do not release the lot, and do not re-integrate or re-run until the failure is investigated and the cause is identified. A system suitability failure is a signal about the method or the instrument as much as about the sample, so the investigation should establish which before any repeat injection is treated as a valid result. Document the failure, the investigation, and the resolution together, because the failed run is part of the lot’s record.
Chiave da asporto: Each answer above stands alone, so you can lift the one you need into a supplier query, an SOP note, or a review meeting without losing its context.
Passaggi successivi
The regulatory outcome is not yours to control; the record set is. Start tomorrow with the phase that is weakest in your current files, usually Phase 4, and work forward. A peptide documentation checklist only helps if someone can hand the file to a reviewer who was not in the room.
If you would like a worked example, MOL Changes supports teams with a sample documentation package covering method files, impurity tables and lot-level release records, which can be used to benchmark your own template before your next audit cycle.
This article is documentation guidance, non consulenza legale o normativa. Consult qualified regulatory counsel before making compliance decisions. Reviewed by the MOL Changes technical documentation team. MOL Changes fornisce sintesi peptidica personalizzata e servizi analitici; this article is educational and does not constitute a product recommendation.