जेनेरिक एलआईएमएस मूल्यांकन पेप्टाइड क्यूए में विफल क्यों होता है
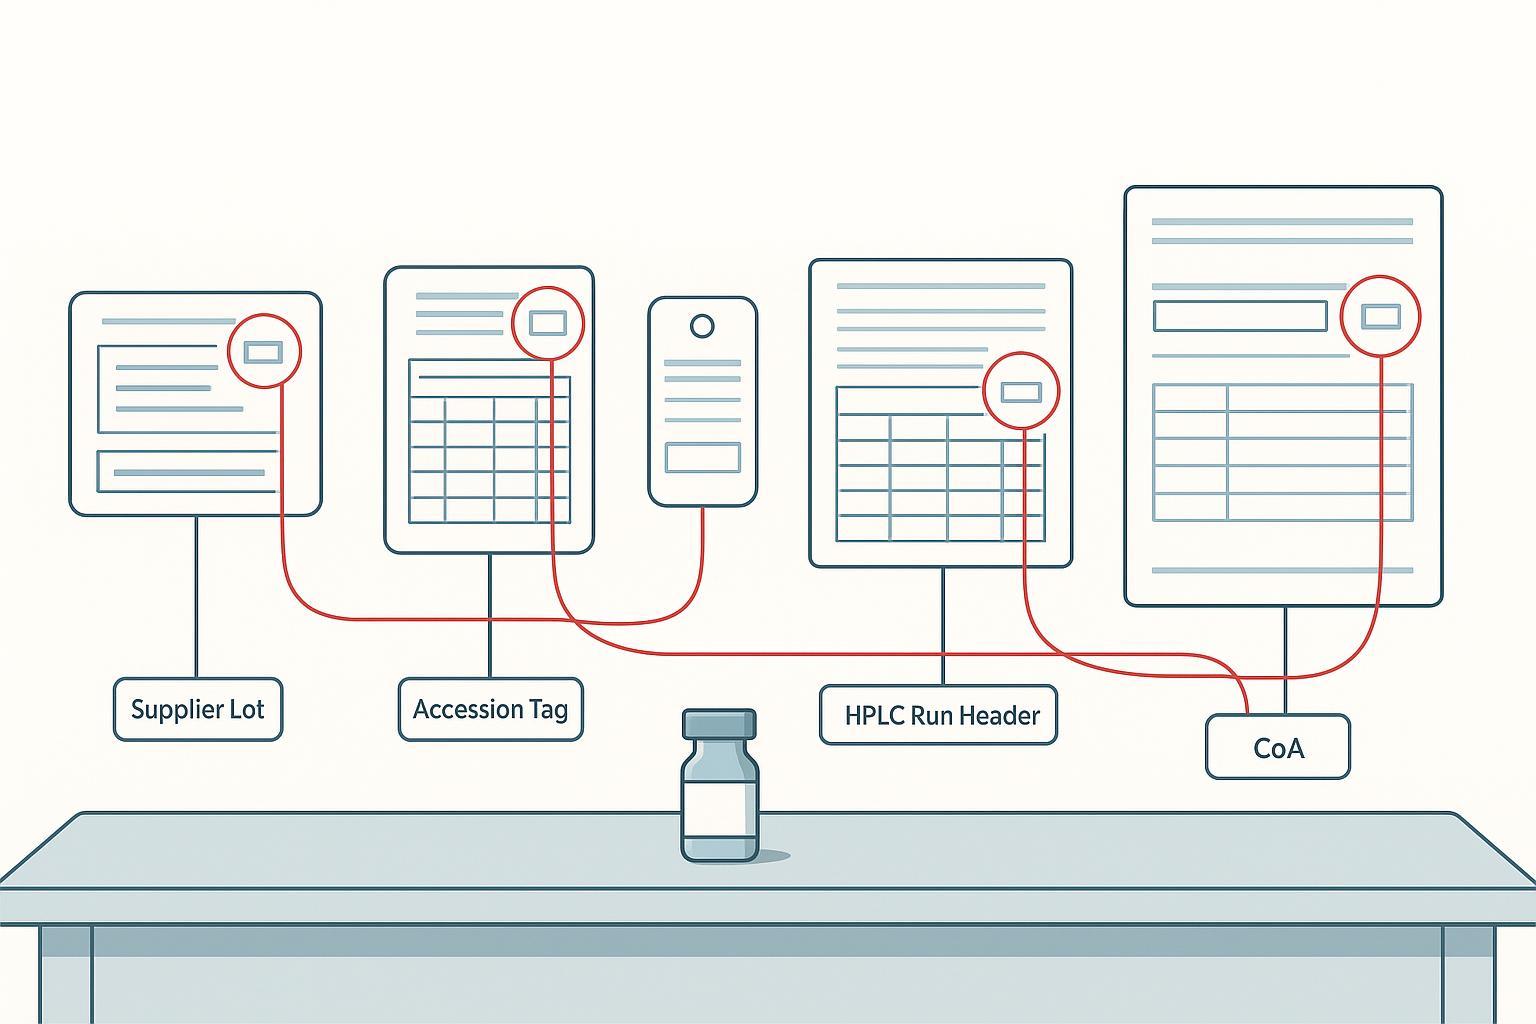
पेप्टाइड लॉट ट्रैसेबिलिटी उस चेकलिस्ट को तोड़ देती है जो अधिकांश प्रयोगशालाओं को जेनेरिक एलआईएमएस गाइड से प्राप्त होती है. पेप्टाइड रिलीज़ पैनल एक नमूने पर एक परीक्षण नहीं है. यह मास स्पेक्ट्रोमेट्री पहचान तक फैला है, उलट-चरण एचपीएलसी शुद्धता, काउंटरियन और ट्राइफ्लूरोएसेटिक एसिड सामग्री, एलएएल परख द्वारा एंडोटॉक्सिन, और बाँझपन, और उनमें से प्रत्येक रिकॉर्ड में ऐसे पहचानकर्ता होते हैं जिनसे बिल्कुल मेल खाना चाहिए: आपूर्तिकर्ता लॉट, आंतरिक लॉट, प्रवेश संख्या, अनुक्रम तालिका, क्रोमैटोग्राम हेडर, और विश्लेषण का प्रमाण पत्र. जेनेरिक गाइड उस समझौते में से किसी का भी परीक्षण नहीं करते हैं.
वे गलत सीमा पर भी रुकते हैं. व्यवहार में, क्रोमैटोग्राफ़िक डेटा प्रबंधन दो प्रणालियों में विभाजित होता है. सीडीएस उपकरण को नियंत्रित करता है, अनुक्रम निष्पादित करता है, क्रोमैटोग्राम को प्राप्त और एकीकृत करता है, सिस्टम उपयुक्तता का मूल्यांकन करता है, और मूल कच्ची फ़ाइल का स्वामी है. LIMS के पास पंजीकरण है, हिरासत की श्रृंखला, परीक्षण असाइनमेंट, बैचिंग, परिणाम एकत्रीकरण, विशिष्टता जांच, रिपोर्टिंग, और स्वभाव रिकॉर्ड. LIMS कार्यसूची को आगे बढ़ाता है और अनुमोदित परिणामों को वापस ले लेता है, जो एक पेप्टाइड मूल्यांकन का हैंडऑफ़ है, उसे मानने के बजाय पूछताछ करनी होती है.
यदि आपकी वर्तमान चेकलिस्ट सामान्य LIMS गाइड से आई है, यह एक डेटा मॉडल के लिए लिखा गया था जिसे कभी भी क्रोमैटोग्राम हेडर के साथ अनुक्रम तालिका का मिलान नहीं करना पड़ता था.
सवाल 1: क्या सिस्टम अनुक्रम और लॉट ट्रैसेबिलिटी को शुरू से अंत तक साबित कर सकता है?
पेप्टाइड लॉट ट्रैसेबिलिटी तभी सिद्ध होती है जब समान पहचानकर्ता अनुक्रम तालिका पर दिखाई देते हैं, रन हेडर और सीओए, और सिस्टम उस श्रृंखला को एक ही क्वेरी में चला सकता है. व्यवहार में, इसका मतलब है कि क्षेत्रों के तीन समूहों को अलग-अलग मॉड्यूल में रहने के बजाय एक-दूसरे से सहमत होना होगा.
|
अनुक्रम तालिका |
हेडर चलाएँ |
संदर्भ मानक |
|---|---|---|
|
अनुक्रम और संशोधन |
डेटा-फ़ाइल नाम और पथ |
मानक नाम और लॉट |
|
नमूना पहचानकर्ता |
उपकरण आईडी |
नियत शक्ति या शुद्धता |
|
पेप्टाइड संश्लेषण विधि का नाम और संस्करण |
समाप्ति या पुनः परीक्षण तिथि |
|
|
अधिग्रहण की तारीख और समय |
अंशांकन वक्र से लिंक करें |
|
|
सेवाएं विश्लेषक आईडी |
||
|
शीशी की स्थिति या कार्यसूची |
साझा पहचानकर्ता नमूना पहचानकर्ता और विधि संस्करण हैं: या तो तोड़ो और श्रृंखला रुक जाती है. इस सूची को अपनी स्वयं की डेमो स्क्रिप्ट की संरचना के लिए व्यवसायी मार्गदर्शन के रूप में मानें, नियामक जाँच सूची के रूप में नहीं.
फिर विक्रेता को इसे पीछे की ओर चलाएं. उन्हें एक सीओए लॉट नंबर दें और उनसे अनुक्रम रिकॉर्ड तैयार करने के लिए कहें, रन, विधि संस्करण और विश्लेषक एक दृश्य में, सिस्टम को छोड़े बिना या हाथ से क्वेरी का पुनर्निर्माण किए बिना.
सवाल 2: एलआईएमएस एलिकोट्स और स्थिरता खींचने के लिए कस्टडी की श्रृंखला को कैसे संभालता है?
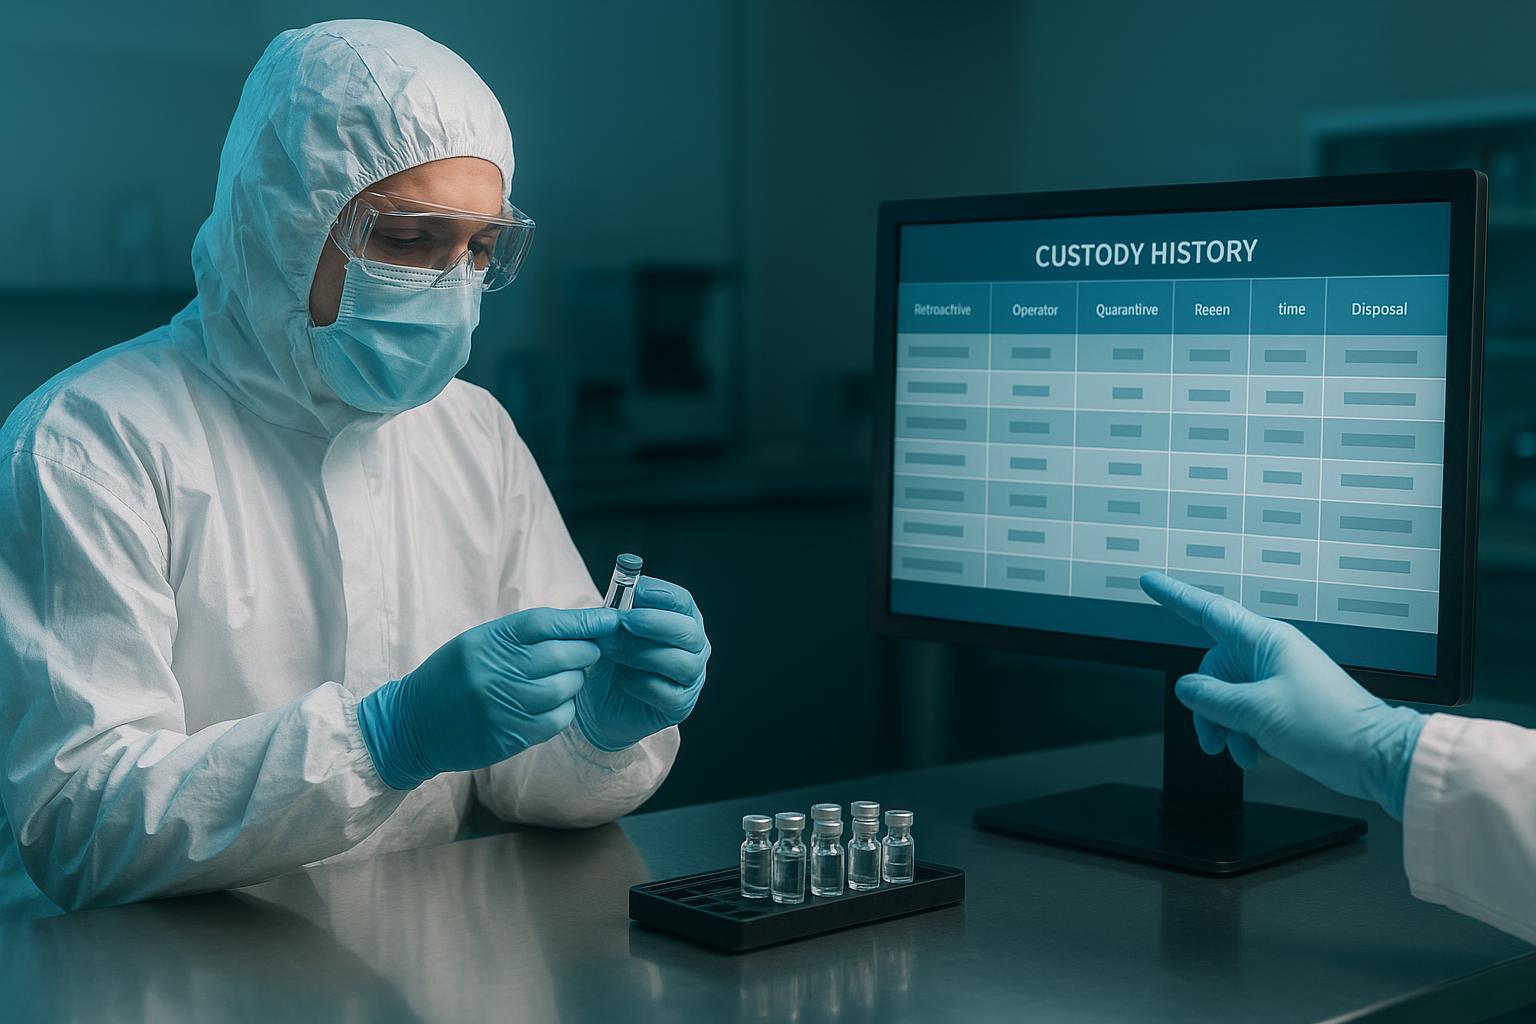
LIMS में हिरासत की श्रृंखला स्थानान्तरण और राज्यों का एक रिकॉर्ड है, हस्ताक्षर ब्लॉक नहीं. प्राप्ति एवं संगरोध, प्रत्येक विभाज्य विभाजन, प्रत्येक जमने/पिघलने की घटना, प्रत्येक स्थिरता पुल बिंदु और अंतिम निपटान प्रविष्टि को अपने स्वयं के टाइमस्टैम्प की आवश्यकता होती है, जिम्मेदार रिकॉर्ड. 21 सीएफआर भाग 11 सुरक्षित की आवश्यकता है, कंप्यूटर जनित, टाइम-स्टैम्प्ड ऑडिट ट्रेल्स जो स्वतंत्र रूप से तारीख को कैप्चर करता है, प्रत्येक क्रिया के पीछे समय और संचालक होता है जो सृजन करता है, इलेक्ट्रॉनिक रिकॉर्ड को संशोधित या हटा देता है. समान विनियमन यह मानता है कि वे रिकॉर्ड एफडीए निरीक्षण के लिए आसानी से उपलब्ध होने चाहिए, यह वह मानक है जिसे हिरासत इतिहास को मैन्युअल पुनर्निर्माण के बिना पूरा करना होता है.
For a peptide lot, that means a reviewer can open one view and see receipt, संगरोध रिहाई, तीन विभाज्य विभाजन, दो स्थिरता खींचती है और एक निपटान प्रविष्टि, each carrying a timestamp, an operator and a reason.
सवाल 3: क्या LIMS कच्चे क्रोमैटोग्राफ़िक डेटा और विधि संस्करणों को नियंत्रित करता है?
Chromatographic data management and method version control decide whether a release decision can be reconstructed years later. The LIMS must know which method version produced each result, and it must block reporting a result against a superseded version. If it cannot resolve that link, the batch record is incomplete no matter how clean the audit trail looks.
The boundary matters here. The chromatography data system owns instrument control, sequence execution, acquisition, integration, system suitability and the native raw file. LIMS के पास पंजीकरण है, custody, परीक्षण असाइनमेंट, बैचिंग, परिणाम एकत्रीकरण, विशिष्टता जांच, reporting and the disposition record. A CDS alone will not carry the lot-level context; a LIMS alone will not hold the raw signal.
Map the method lifecycle to a recognised framework. मैं Q2(आर2), the harmonised analytical procedure validation framework issued in March 2024, covers validation principles including the analytical use of spectroscopic data, and supports justified post-approval change management. For the instruments themselves, USP 〈1058〉 frames instrument qualification as fitness for intended use, official as of 1 August 2017.
In the demo, ask the vendor to revise a method, then show what happens to results already reported under the prior version.
सवाल 4: विशिष्टता से बाहर की जांचों को कैसे रूट और संरक्षित किया जाता है?
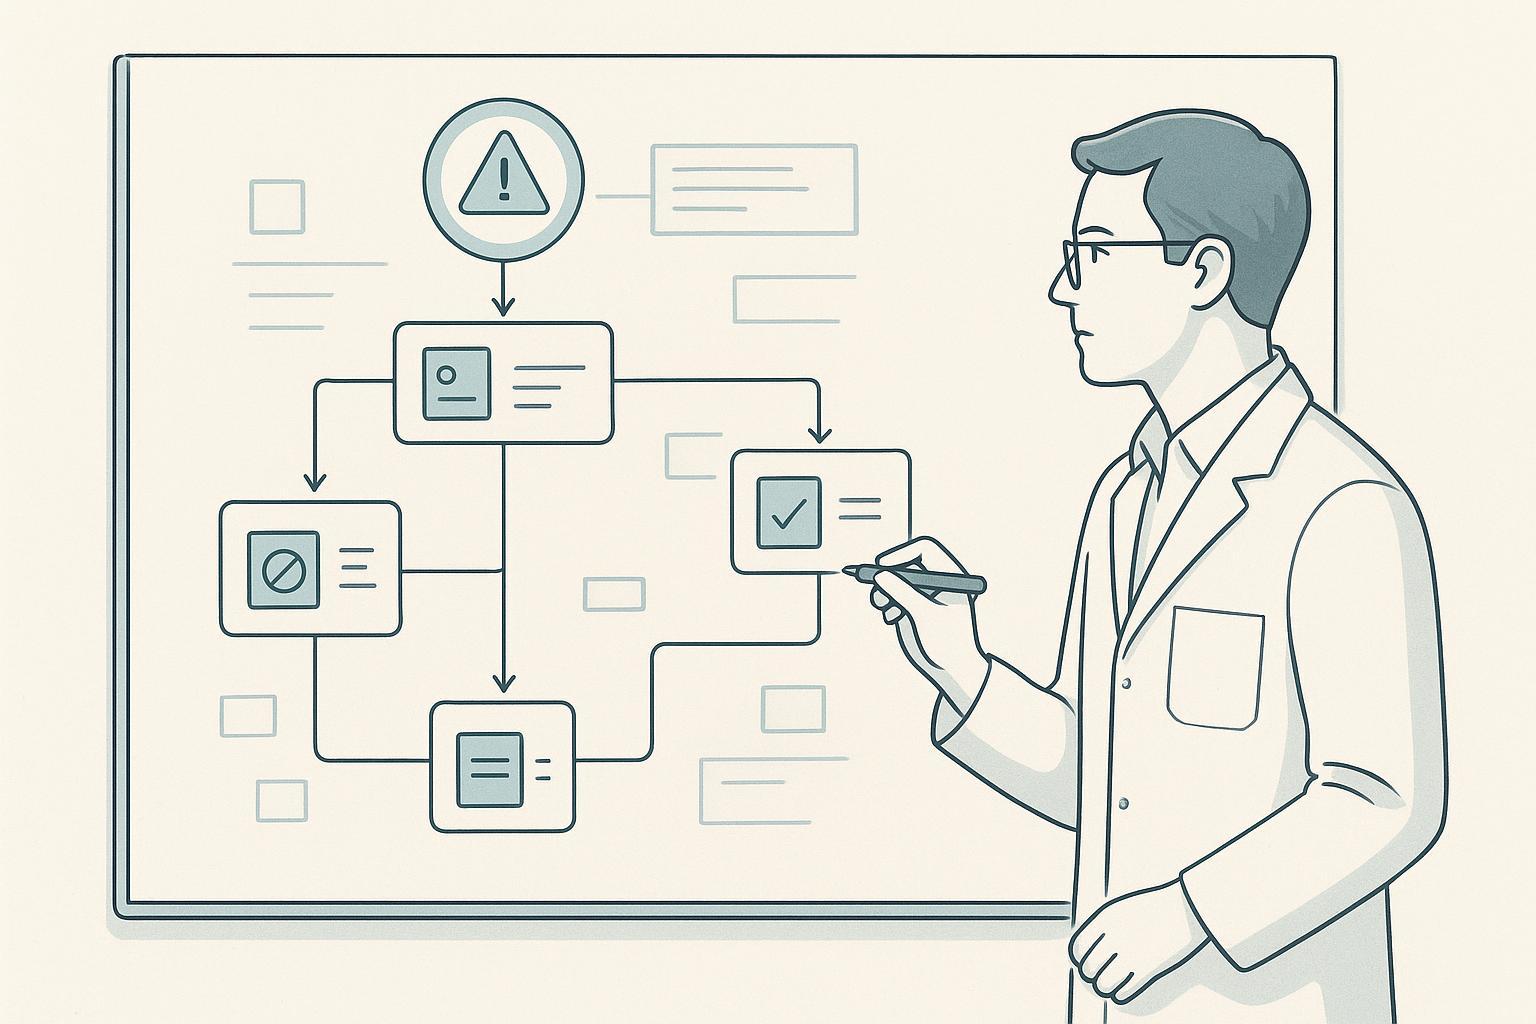
An out-of-specification investigation in a peptide QC lab is a routing and preservation problem before it is a compliance exercise. Industry-standard two-phase practice, framed by the FDA’s guidance on investigating out-of-specification test results, starts with a Phase I laboratory investigation: the lab checks its own work for an assignable cause before the result is treated as a product failure. Only when Phase I finds no assignable laboratory cause does the event escalate to Phase II, where the investigation moves beyond the laboratory.
The routing matters less than what survives it. For any OOS event, the record an auditor must be able to reconstruct includes the original reported result with sample ID, तरीका, instrument, run sequence, date and time, analyst, and batch or lot context; the raw data behind it, including chromatograms, integrations, weights, calculations, and system-suitability and calibration records; an audit trail of every addition, विलोपन, परिवर्तन, reprocessing, and re-integration, with who authorised each one and why; the investigation record itself, with its phase assignment, evidence reviewed, conclusions, कैपा, and disposition; and the full retest history showing why retests were permitted, whether the initial result was invalidated, and which result became reportable (NCI Frederick, Managing Out-of-Specification Test Results or Unexpected Test Results, 2022).
A LIMS that stores only the final disposition cannot reconstruct any of this. Ask the vendor to walk one OOS event through the system end to end, from the first reported value to the reportable result.
सवाल 5: क्या विश्लेषण का प्रमाणपत्र केवल अनुमोदित परिणामों से उत्पन्न होता है?
Certificate of analysis generation is a controlled-document function, not a reporting template. A CoA is assembled from approved results, and its header fields must agree exactly with the sequence table, run header and reference standard records for that lot. The fields that carry the most weight are product name, बहुत संख्या, COA date and testing laboratory, because those four are what a customer checks first and what an auditor reconciles against the batch record.
That is why the release-blocking test belongs in the demo. Ask the vendor to generate a CoA for a lot that still has an open out-of-specification investigation against it, then watch what the system does. A LIMS built for peptide QA refuses, flags, or routes for approval. A reporting tool prints the certificate anyway, and you have just seen how a non-conforming lot reaches a customer.
|
CoA header field |
Source record it must agree with |
|---|---|
|
Product name |
अनुक्रम तालिका |
|
Lot number |
Sequence table and run header |
|
COA date |
Approval record for the released results |
|
Testing laboratory |
Run header and reference standard records |
If the vendor cannot show that agreement in the demo, treat the CoA as unverified until they do.
एक रक्षात्मक साक्ष्य श्रृंखला शुरू से आखिर तक कैसी दिखती है
A defensible peptide lot traceability chain is a sequence of handoffs, and each handoff needs a named record plus a control that keeps the record honest. Walk it in the demo from the vial label through accession, analytical run, method version, analyst, समीक्षा, disposition, और विश्लेषण का प्रमाण पत्र. At every one of those steps, ask which record carries the handoff and what stops that record from being edited after the fact. Chain of custody in the LIMS is only as strong as its weakest handoff, so a system that names the record but cannot show the control is showing you a form, नियंत्रण नहीं.
MOL Changes supports this kind of staged handoff, and can be used to keep method versions and review records attached to the lot they belong to.
पेप्टाइड क्यूए के लिए एलआईएमएस चयन में सामान्य गलतियाँ
The most common failure in choosing a LIMS for peptide QA is evaluating on a feature checklist instead of asking the vendor to produce a record. Checklists are easy to score and hard सिंथेटिक पेप्टाइड्स to falsify, which is exactly why they survive procurement. The five mistakes below are the ones that cost labs the most, and each has a cheap fix you can apply in the next demo.
Accepting a demo on synthetic data. Vendors demo with clean, pre-built lots because real data exposes gaps. Ask them to start from one of your own CoA lot numbers and walk to the sequence record, रन, the method version and the analyst who signed it. If the walk breaks anywhere, you have found your answer.
Treating the CDS/LIMS boundary as an integration detail. It is a data-ownership decision. Whoever holds the raw chromatographic data controls the audit trail, so settle that before you discuss connectors.
Letting method versioning be a free-text field. Free text cannot prove which method produced a result. Require a version history that binds each result to the method revision in force at the time of the run.
Assuming CoA generation is a reporting feature. It is a release control. Ask what the system does when an out-of-specification investigation is still open at the moment someone tries to issue the certificate.
Not asking what happens to an open OOS at release. This is the question that separates a defensible system from a document repository, and it takes thirty seconds to ask.
उत्तर कैसे प्राप्त करें और निर्णय कैसे लें
Score each of the five questions on three axes, and score the vendor’s behavior, not the feature list. पहला: did they demonstrate the record in the system, or only describe it? A traceability claim shown live, with a real lot walked from synthesis through release, is worth more than a slide. दूसरा: was the evidence exportable? Ask them to export the audit trail, the method-version history, or the OOS record as a file you can open. If the answer lives only inside their interface, you cannot audit it. तीसरा: did the answer survive a follow-up? Ask what happens when a stability pull is split across two analysts, or when a method is revised mid-study. Vendors who know their product answer immediately; vendors who do not will offer a call with an engineer.
For a LIMS for peptide QA, the scoring criteria should also include the cost and timeline of validation, because that is often the deciding factor. Regulated LIMS validation is reported at roughly $15,000 को $80,000 for validation alone, with full regulated implementations commonly reported at $150,000 को $500,000 and timelines of 6 को 12 months single-site and 18 को 36 months multi-site (1LIMS, पुनर्प्राप्त 2026-08-10). Treat these as indicative only: they come from vendor and consultancy estimates that vary widely, with no stated methodology or sample, so they should not anchor your decision. Use them to frame a budget conversation, not to choose a system.
|
सवाल |
Demonstrated the record |
Evidence exportable |
Survived follow-up |
|---|---|---|---|
|
1. Sequence and lot traceability |
|||
|
2. Chain of custody for aliquots and stability pulls |
|||
|
3. Raw chromatographic data and method versions |
|||
|
4. OOS investigation routing and preservation के बारे में |
|||
|
5. CoA generated from approved results only |
Two limitations deserve a place in the scoring. Instrument-vendor lock-in is real: a system built tightly around one chromatography vendor’s data format can make a second instrument line expensive to onboard later. Migration cost is the other. Moving historical peptide data out of a legacy system, with its audit trails intact, is a project in itself, and vendors rarely quote it upfront. Ask both questions before you sign, and score the answers honestly.
If the scoring leaves you with two viable options, talk to a peptide QC specialist who has run these systems in a regulated lab, not just a sales engineer.
अक्सर पूछे जाने वाले प्रश्नों
क्या क्रोमैटोग्राफी डेटा सिस्टम अकेले पेप्टाइड क्यूए ट्रैसेबिलिटी को कवर कर सकता है?
नहीं. A chromatography data system is the system of record for acquisition, integration, and the native raw file, but registration, custody, बैचिंग, विशिष्टता जांच, and the disposition record sit in the LIMS. That boundary matters for your evaluation because chromatographic data management and peptide lot traceability are two different record chains, and only one of them ends in a release decision. Ask the vendor to show where the CDS record is referenced by the LIMS record, and what happens to that reference when a result is reprocessed.
क्या होगा यदि विक्रेता डेमो में विधि-संस्करण इतिहास नहीं दिखा सकता है?
Treat it as an unresolved release-blocking control, not a roadmap item. Method version control is what lets you prove which procedure produced a released result, so a demo that cannot display it is an incomplete demo, not a scheduling problem. Ask for the method lifecycle in writing, including how a version is retired and how historical results stay bound to the version that generated them, then ask for a reference lab that has been audited on exactly that. If the vendor cannot supply either, the out-of-specification investigation path downstream is unverifiable too.
पेप्टाइड QC LIMS को कितने सत्यापन की आवश्यकता है??
Scope validation to the records your lab releases on, and map instrument qualification to USP 〈1058〉 “fitness for intended use” rather than treating it as a separate exercise. In practice that means the validated scope covers registration, custody transfer, specification evaluation, और स्वभाव रिकॉर्ड, with the instruments feeding those records qualified for the purpose they serve. A LIMS for peptide QA does not need to be validated as a whole system beyond what those release decisions depend on.
क्या कोई ईएलएन इन पांच प्रश्नों में से किसी को बदलता है?
नहीं. An ELN changes where the analyst writes, not who owns custody, method version, or the disposition record. If an ELN is in the stack, the same five questions apply to whichever system holds those records, and you should confirm in the demo which one that is.
निष्कर्ष
You now have a working evaluation framework: five questions for a LIMS for peptide QA, each paired with a specific record to demand and a specific request to send. Sequence and lot traceability maps to the full analytical record set, which for a typical peptide lot runs to roughly six to eight core documents covering MS identity, RP-HPLC purity, प्रतिवाद, अन्तर्जीवविष, बांझपन, और अवशिष्ट विलायक (Polypeptide QC specifications, undated). Chain of custody maps to aliquot and stability-pull logs. Raw chromatographic data and method versions map to the instrument export and version history. OOS routing maps to the investigation record. CoA generation maps to the approved-results gate.
The value of asking this way is that it moves the conversation from a feature list to an audit. A vendor can claim traceability; only the records show whether it holds.
The next step is deliberately small. Request a representative CoA and analytical data package, then run the five questions against it before you book a second demo. If the answers are thin, ask to speak with a peptide QC specialist on the vendor’s team and put the same five questions to them.
खुलासा: MOL Changes has a commercial interest in peptide quality standards and LIMS evaluation practice.