Azịza ngwa ngwa
Lonza's integrated DNA-to-IND offering is built for recombinant formats. Its program page names monoclonal antibodies, bispecific antibodies and Fc-fusion prot…
The integrated model is built for recombinant molecules, not for custom peptides in DNA-to-IND programs
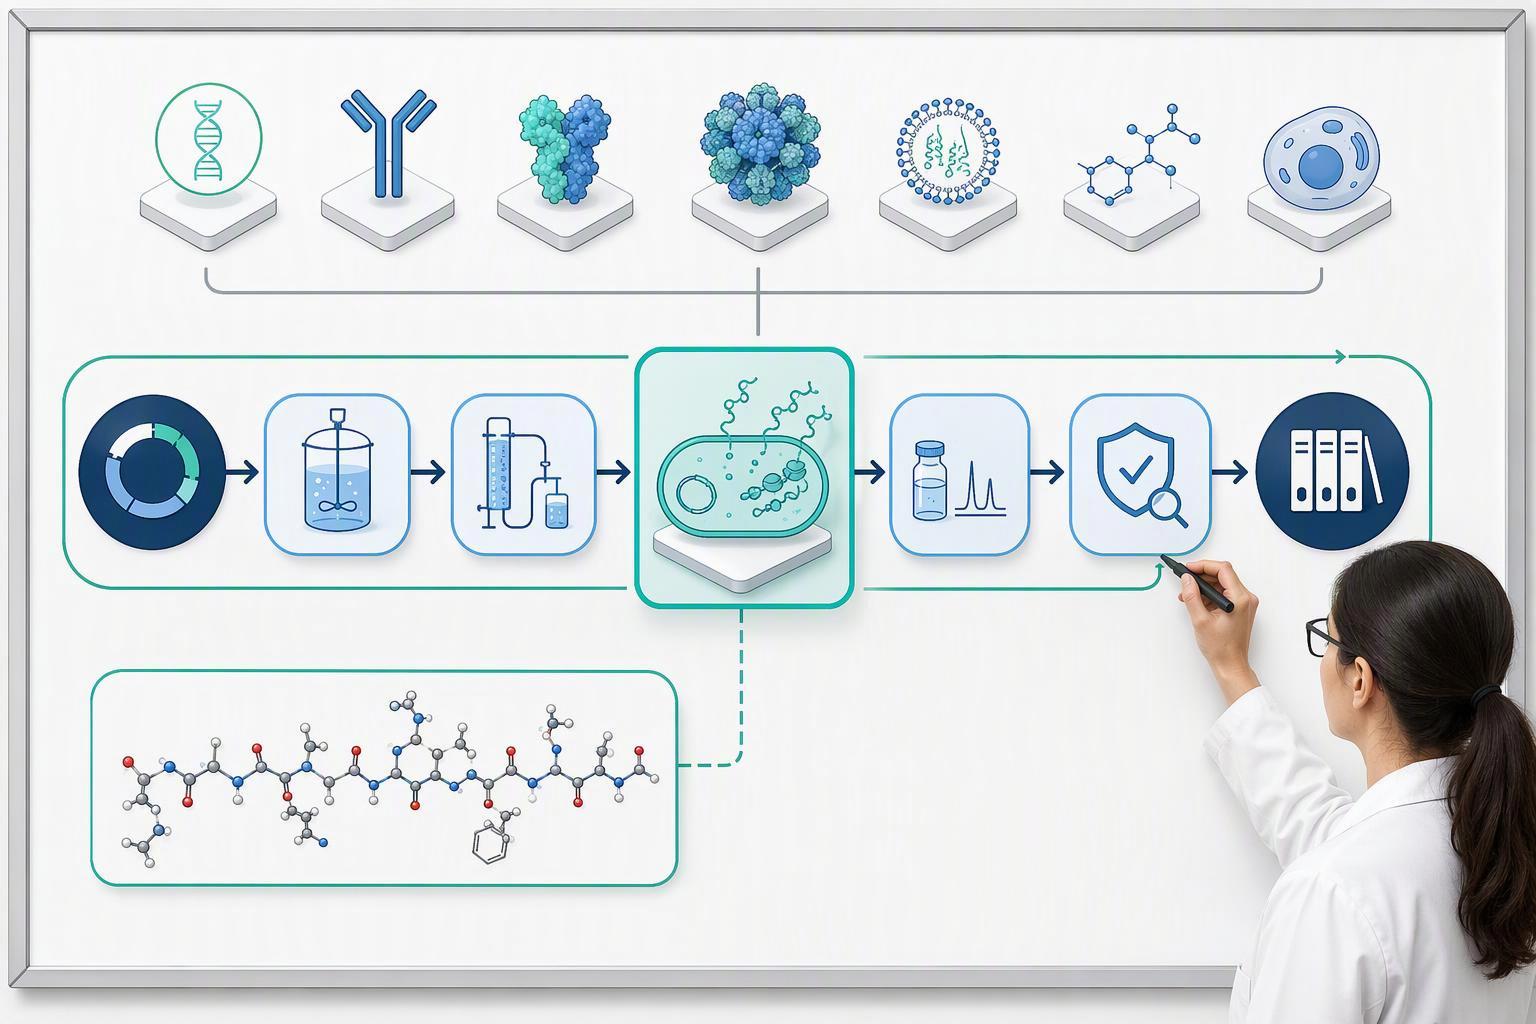
The integrated pathway is defined by recombinant expression, and that definition, not a gap in service coverage, is what places synthetic peptides outside it. Lonza’s program page describes one coordinated CMC workflow that brings cell line development, process and formulation development, analytical testing and regulatory documentation together by “minimizing handovers and aligning all CMC activities” (Lonza DNA-to-IND program page, emechiri emechi, ewepụtara 2026). Every step in that chain begins with a transfected cell.
The October 2026 expansion extended the same chain to non-mAb biologics, adding bispecific antibodies, Fc-fusion proteins and high-concentration drug product formulations, and moving high-concentration formulation development earlier in the process (Ndụmọdụ mgbasa ozi Lonza). The molecule classes named are all recombinant formats.
The programme’s defined entry points run from vector construction and manufacturability assessment through DNA to tox drug substance or drug product batch, DNA to research, master or production cell bank, transfer of an established cell line or process, and bulk drug substance or purified product, with the option to run fully integrated to IND or pause at defined milestones (Lonza DNA-to-IND program page, emechiri emechi, ewepụtara 2026). A chemically synthesized peptide has no vector, no cell line and no transfection step to enter at.
The vendor-stated timelines carry the same frame. The milestone table lists six months from DNA sequence to IND/IMPD, DNA to tox in two months and early de-risking in under 11 izu, with a footnote limiting the claim to “from DNA transfection to delivery of IND-enabling CMC modules. Usoro iheomume a na-ahụkarị. Subject to terms and conditions” (Lonza DNA-to-IND program page, emechiri emechi, ewepụtara 2026). The clock starts at transfection. For a wholly chemically synthesized peptide, DNA transfection is not the relevant start milestone.
The two frameworks draw the peptide boundary differently, and the difference is chemistry
Production route, product characteristics, indication and jurisdiction influence the regulatory pathway. Post-chain chemistry is one consideration, not the single variable deciding which framework applies.
The two regimes then diverge on where a modified peptide lands. The EMA guideline accepts protected natural L-amino acid derivatives, and in justified cases short peptide fragments, as starting materials, but states that polypeptide fragments undergoing only limited further GMP modification, such as cyclisation or conjugation to non-peptide moieties like PEG chains, bụ ndị etiti kama ịmalite ihe. It also treats an activated PEG derivative such as an NHS ester as an intermediate itself (Ntuziaka EMA, dị irè 1 June 2026).
FDA withdrew the 2021 guidance in July 2026. Modification or lipidation alone does not universally exclude a generic-drug pathway; assess sameness to the reference product, current product-specific guidance and the actual application requirements. FDA’s withdrawal and revised-draft update.
The EMA guideline addresses relevant synthetic-peptide and conjugation quality questions. FDA pathway eligibility must be assessed separately for the proposed product and reference drug; the amount of added chemistry does not create a common EMA/FDA exclusion rule.
What the impurity and specification record actually requires of a peptide
A peptide’s specification is written from peptide-specific impurity classes and thresholds, not inherited from either adjacent framework. The categories the EMA guideline names are stereoisomers, deletion sequence, truncated sequence and insertion sequence (EMA guideline landing page, dị irè 1 June 2026). The thresholds the EMA points to come from Ph. Euro. monograph izugbe 2034: report an impurity above 0.1%, mata n'elu 0.5%, tozuo oke 1.0%, with no universal numerical limit for unspecified or total peptide-related impurities, which are set product-specifically and justified case by case (Ntuziaka EMA, dị irè 1 June 2026).
Peptide N'ogbe The thresholds that do not apply are worth stating plainly, because they are a common source of error. I Q3A(R2) sets reporting at 0.05%, njirimara na 0.10% ma ọ bụ 1.0 mg/day and qualification at 0.15% ma ọ bụ 1.0 mg/day for a maximum daily dose at or below 2 g/ụbọchị (I Q3A(R2), nlegharị anya ugbu a). Those numbers are not peptide limits, and the EMA guideline states directly that the Q3A limits “are not applicable” to synthetic peptides.
Conjugation adds attributes that a chain-only specification misses. The EMA guideline requires control of the free unconjugated peptide and the free form of the conjugate moiety, dị ka PEG efu ma ọ bụ njikọ njikọ, and states that di-PEGylation, multi-PEGylation and non-site-specific PEGylation products should be adequately controlled (Ntuziaka EMA, dị irè 1 June 2026). It also requires that all steps of the intermediate synthesis from the defined starting material be performed under GMP, with full process information in Module 3 Section 3.2.S.2.2.
The specification guideline that excludes synthetic peptides is not the frame to reach for. ICH Q6B defines a specification as a list of tests, references to analytical procedures and acceptance criteria, and its impurity taxonomy is built for recombinant expression: cell substrate-derived, cell culture-derived and downstream-derived (I Q6B, Nzọụkwụ 4, 1999). That taxonomy does not transfer to a solid-phase synthesis, where the impurities are enantiomeric and diastereomeric species, amino asịd na-enweghị nchebe, dipeptides na beta-alanyl adịghị (Ntuziaka EMA, dị irè 1 June 2026).
Where a specialist adds value: feasibility triage before the route is fixed
The case for an early feasibility check is arithmetic, not preference. Thirty couplings at 99% effective yield give 0.99³⁰, ike ike 74% cumulative retention before deprotection, ikpochapu, purification and isolation losses (Pharma’s Almanac). A sequence that couples poorly at three positions loses more than the model predicts, and the loss compounds silently until the crude profile is measured.
That is why route selection is a sequence-specific decision rather than a platform default. A 2025 review of therapeutic peptide discovery and synthesis reports that chemical synthesis suits peptides of 50 residues or fewer, where truncated sequences, racemisation and side reactions require HPLC purification, while biological synthesis suits longer sequences but introduces Peptide 1 Onye nrụpụta host-cell proteins, endotoxins and misfolding (Zheng et al., Int. J. Mol. Sci., 2025). The same review notes that minor deviations in amino acid sequence, post-translational modification or impurity profile can critically affect pharmacological activity, pharmacokinetics and immunogenicity.
Scale changes the control points, not just the batch size. Custom peptide synthesis spans milligrams for discovery and assay standards, grams for process development, toxicology and early clinical work, tens to hundreds of grams or around a kilogram per batch for clinical and niche products, and kilograms to multi-kilograms per batch for commercial API, with sequence, modification and purity requirements mattering as much as nominal batch size (Pharma’s Almanac, 18 Mee 2026). Njikọta zuru oke, nchedo na cleavage omume, crude adịghị ọcha ibu, preparative purification loadability and counter-ion or salt form are the points that decide whether a campaign scales, and they are cheapest to test before the route is locked.
Emegharịrị, labeled and conjugated peptides are where the integrated model stops
<klas ọgụgụ ="blọọgụ-akara ngosi" ụdị ="oke-obosara:640px;oke:1.5rem akpaaka;font-nha:12px;ahịrị-ịdị elu:1.3;agba:eketa"> <svg viewBox="0 0 560 380" ụdị ="oke-obosara: 100%; elu: akpaaka; font-ezinụlọ: 'Inter', usoro-ui, sans-serif; agba: #1e293b; --chaatị-mechiri emechi: #64748b;" ọrụ ="img" aria-labeledby="2025-peptide-and-oligonucleotide-cdmo-revenue-split-title 2025-peptide-and-oligonucleotide-cdmo-revenue-split-desc"> <ịke> /* Onye nrụpụta na-etinye eserese ahụ n'ime iframe nke ihe ndabere ya dị oji, na iframe bụ ọdịdị dị iche na chaatị ahụ, ya mere eriri igbe akwụkwọ ozi gosipụtara dị ka okpokolo agba gbara ya gburugburu (prod 09-22). Akwụkwọ dị n'ime iframe pụtara ìhè na ndabara; esere ya ebe a na-ekpuchi ndabere elementrị, na mgbe e debere chaatị ahụ kama nke a dabara na ihe ọ bụla. Debe brackets n'akuku na ngọngọ a: A na-atụgharị akụkụ ụdị n'ime SVG dị ka ọdịnaya nke mba ọzọ, ya mere otu nkwado kpafuru akpafu na-atụfu iwu niile dị na ya. */ html, ahu { ndabere: #fffff; oke: 0; } @mgbasa ozi (na-ahọrọ-ebelata-mmegharị: belata) { * { animation: onweghi !mkpa; mgbanwe: onweghi !mkpa; } } </ịke> <ziri ezi x="0" y="0" obosara ="560" elu ="380" rx="8" juru ="#fffff" /> <aha id="2025-peptide-and-oligonucleotide-cdmo-revenue-split-title">2025 peptide and oligonucleotide CDMO revenue split</aha> <desc id ="2025-peptide-and-oligonucleotide-cdmo-revenue-split-desc">peptide and oligonucleotide CDMO revenue split: values for Peptides, Oligonucleotides. 2025 peptide and oligonucleotide CDMO revenue split. data donut: Peptides 66.7; Oligonucleotides 33.3.Source: Nchọpụta Nleba anya, Peptide & Oligonucleotide CDMO Market 2026.</desc> <ederede x ="280.0" y="29" ederede-arịlịka ="etiti" font-size ="18" font-arọ="800" juru ="Agba ugbu a">2025 peptide and oligonucleotide CDMO revenue split</ederede> <path d="M 238.00 93.00 A 92 92 0 1 1 158.23 230.83 L 192.91 210.91 A 52 52 0 1 0 238.00 133.00 Z" juru ="#f97316" /> <path d="M 158.23 230.83 A 92 92 0 0 1 238.00 93.00 L 238.00 133.00 A 52 52 0 0 0 192.91 210.91 Z" juru ="#38bdf8" /> <ederede x ="238" y="182" ederede-arịlịka ="etiti" font-size ="22" font-arọ="800" juru ="Agba ugbu a">100</ederede> <ederede x ="238" y="201" ederede-arịlịka ="etiti" font-size ="10" juru ="anyi(--chaatị-mechiri emechi, Agba ugbu a)">ngụkọta</ederede> <ziri ezi x="360" y="96" obosara ="11" elu ="11" juru ="#f97316" /> <ederede x ="378" y="105" font-size ="11" juru ="Agba ugbu a">Peptides 66.7</ederede> <ziri ezi x="360" y="124" obosara ="11" elu ="11" juru ="#38bdf8" /> <ederede x ="378" y="133" font-size ="11" juru ="Agba ugbu a">Oligonucleotides 33.3</ederede> <ederede x ="280.0" y="366" ederede-arịlịka ="etiti" font-size ="10" juru ="anyi(--chaatị-mechiri emechi, Agba ugbu a)">Isi mmalite: Nchọpụta Nleba anya, Peptide & Oligonucleotide CDMO Market (2026)</ederede> </svg> </ọgụgụ>
The starting-material logic is where the two regimes separate most sharply. The EMA guideline requires the choice of starting material for the conjugation component to be justified under ICH Q11, and states that activation of a suitable PEG starting material is part of the manufacturing process, so an activated PEG derivative such as an NHS ester may be considered an intermediate rather than a starting material (Ntuziaka EMA, dị irè 1 June 2026). It also expects separate documentation for each supplier of the conjugation moiety or linker, and a compiled specification elaborated by the peptide-conjugate manufacturer.
The Synthesis Of Aspirin The practical consequence is that a program adding labels, njikọ, PEG chains or lipid tails is running a chemistry project alongside a peptide project. Nke 2025 review’s comparison table makes the split explicit: chemical synthesis enables non-natural amino acids, isotopic labelling and site-specific modifications, while biological synthesis is limited to natural amino acids unless genetic or enzymatic engineering is added (Zheng et al., Int. J. Mol. Sci., 2025). That flexibility is exactly what a recombinant-expression platform cannot absorb. Peptide 2 Onye nrụpụta
The handoff is a specification and method-transfer problem, not a timeline problem
When a peptide program moves between partners, the transferable asset is the specification and the validated methods behind it, and the governing validation guideline is shared with the biologic route. I Q2(R2), nakweere 1 Nọvemba 2023 with an error correction on 30 Nọvemba 2023, applies to analytical procedures used for release and stability testing of drug substances and products, covering assay, ike, ịdị ọcha, impurity as a quantitative or limit test, na njirimara, and can be applied phase-appropriately during clinical development (I Q2(R2), nakweere 1 Nọvemba 2023). Its scope is defined by use case rather than molecule type.
The peptide-specific side of the package is less standardised. The EMA guideline expects full information in Module 3 Section 3.2.S.2.2, including flowchart, process description with all process steps, raw materials and manufacturing process controls, and requires all intermediate synthesis steps from the defined starting material to be performed under GMP (Ntuziaka EMA, dị irè 1 June 2026). For a conjugated peptide, that package also carries the conjugation chemistry description and the supplier documentation for each conjugation moiety or linker.
Microbial and endotoxin control are calculated, not fixed. The endotoxin limit is derived as K/M, ebe K 5 EU/kg/hour for most parenteral routes, 0.2 EU/kg/hour for intrathecal and 100 EU/m²/hour for body-surface-area dosing, with M the maximum recommended human dose per kg per hour (USP General Isi <85>, isi ugbu a, ewepụtara 2026). Nnwale ịmụ nwa n'okpuru USP <71> is qualitative: Custom Pna Synthesis it detects viable microbial contamination and does not demonstrate absolute sterility or quantify contamination (USP <71> Nnwale ịmụ nwa, harmonisation text). Neither limit travels as a number in a transfer document; both travel as methods and calculations.
The evidence base does not support a benchmark for transfer failure or delay. No primary or peer-reviewed figure for technology-transfer failure rates was located, so the risk is best argued from the regulatory mechanism: the specification, the validated methods and the GMP process description are what a receiving site must reproduce, and their completeness determines how much of the program is rebuilt.
A decision rule for partnering reviews
The rule below turns the regulatory boundary into a partnering test. It is written as an if/then list so it can be applied directly in a vendor review.
If the candidate is a chemically synthesized peptide of roughly 50 residues or fewer with no post-chain modification, then a specialist synthesis and analytics partner is the fit, and an integrated recombinant pathway is the wrong entry point.
For a modified construct, identify the relevant conjugation chemistry, site specificity, starting materials and likely impurities. Controls for free PEG or multiple PEG attachments apply when PEG chemistry is involved, not automatically to every isotope label or noncanonical residue.
If the sequence has known difficult segments, then run the feasibility and route check before the route is fixed, n'ihi na 30 couplings at 99% per-step efficiency retain only about 74% of the chain before purification (Pharma’s Almanac, 18 Mee 2026).
If the program is a recombinant biologic, then the integrated model is the right frame, and the peptide-specific thresholds and conjugation requirements in this article do not apply.
The market context supports treating peptide programs as a distinct buying decision rather than a subset of biologics. The peptide and oligonucleotide CDMO market was USD 3.1 ijeri 2025 and is forecast at USD 8.1 ijeri site 2033 na a 12.9% CAGR, with peptides holding 66.7% nke 2025 revenue (Nchọpụta Nleba anya, 2026 mbipụta, 2025 afọ ntọala). Both figures come from a report vendor, and the CDMO number covers peptides and oligonucleotides combined. The wider peptide therapeutics market was USD 140.9 ijeri 2025, forecast at USD 294.6 ijeri site 2033 na a 8.7% CAGR (Nchọpụta Nleba anya, 2026 mbipụta, 2025 afọ ntọala). The biologics CDMO market the integrated model sits in is projected at USD 62 billion with an 11.7% CAGR (GlobalData via GlobeNewswire, 14 October 2026). All three are report-vendor models. Oligopeptide Synthesis
Ajụjụ a na-ajụkarị
Do the timelines Lonza announced apply to a peptide program?
Mba, because the clock those timelines measure starts at DNA transfection, and the formats named in the same material are recombinant: monoclonal antibodies, bispecifics and Fc-fusion proteins (Lonza DNA-to-IND program page, emechiri emechi, ewepụtara 2026). A chemically synthesized peptide never passes through that step, so the schedule is not a benchmark a peptide team can borrow. What transfers between the two is the regulatory destination, not the route or the calendar.
Can a specialist carry a program to IND without an integrated provider?
Ee. The Module 3 na Q2(R2) obligations do not change with the size of the partner, so the difference is not what gets filed but who holds the specification and the validated method set at filing. A specialist can hold both, provided the discovery to development peptide handoff is documented as a controlled transfer rather than an informal one. What an integrated provider adds is convenience of coordination, not a regulatory category the specialist cannot occupy.
What if a program is already partway down an integrated path?
Transfer the specification, the methods and the comparative batches instead of restarting. Restarting discards the analytical history that supports the specification, and rebuilding it costs more than moving it. The transfer package is the unit of work: product-specific limits, validated release and stability methods, and the process description that shows where the defined starting material sits.
Mmechi
A peptide program does not inherit the small-molecule impurity framework, and it does not inherit the biologic one. I Q3A(R2) excludes peptides, ICH Q6B excludes synthetic peptides, and the EMA guideline effective 1 June 2026 supplies the peptide-specific framework in their place, with Ph. Euro. 2034 thresholds and product-specific limits. That is the structural fact behind the partnering question, and it is independent of any vendor’s timeline.
The integrated DNA-to-IND model is a coherent answer to a recombinant problem. Its named molecule classes are mAbs, bispecifics and Fc-fusion proteins, its entry points begin at vector construction, and its clock starts at DNA transfection. A chemically synthesized peptide has none of those inputs, so the fit question is answered before cost or speed enters the discussion.
The next step is a feasibility assessment on the specific sequence, run before the route is fixed. Sequence-level coupling risk, mgbanwe chemistry, scale target and the applicable specification framework determine whether a specialist partner is the right choice, and the arithmetic of per-step yield makes that assessment cheapest at the earliest stage.