When to run this review, and what to have ready
With the standard now published, the transition clock is already the practical question. ANSI’s release marking the first full revision in a decade states that the transition period is still “in the approval process” and will be set by Global ACI, with a three-year window expected rather than settled. Two consequences follow. SGS’s published-revision bulletin confirms that existing ISO 9001:2015 certificates remain valid during the transition, and that transition audits must be planned with your certification body ahead of the deadline. Ayrı ayrı, accredited certification bodies must themselves be transitioned by their accreditation body before they can issue 2026 certificates, so your chosen body’s readiness is worth confirming early.
|
What to have ready |
Who should be in the room |
Dated milestone |
|---|---|---|
|
Current QMS document set |
QA lead |
16 Eylül 2026: standard published (ANSI) |
|
Supplier list with risk tiers |
Kalite kontrol lideri |
Transition window expected at three years, not yet final (ANSI) |
|
One live lot’s full record packet |
Production lead |
12-16 Oct 2026: ISO/TC 176 plenary, London (ISO/TC 176) |
|
Deviation log with CAPA links |
Supply chain lead |
|
|
Client specification versions |
Client-facing technical lead |
The revision touches every clause from 3 ile 10 and adds a new Annex A, başına BSI’s clause-by-clause summary of the revision, while keeping the Annex SL structure. ISO/TC 176’s own release announcement, gönderildi 23 Eylül 2026, links both “What’s Changing?” and “Transition Guidance” for readers who want the primary text.
Six-box review sequence, with the evidence artifact named under each box.
One scope note before you start: the review below is derived from the six areas a peptide quality system must control, not from a clause-by-clause reading of the standard. Treat it as a prioritisation aid, not a gap assessment.
Kontrolü değiştir: does your system cover communication and effectiveness, not just approval?
A change review that stops at approval does not cover clause 6.3 as written. A clause-by-clause walkthrough of the 2026 text reports that “Planning of changes” now also expects availability of information, how the change is communicated, how effectiveness is monitored and evaluated, and how results are reviewed (isomanaged knowledge base, geri alındı 2026-09). The ICH Q10 change management guideline points the same way, describing change management built on quality risk management with regulatory filing impact assessed and post-implementation verification that objectives were met. Treat the Q10 extension as unconfirmed until the ICH PDF is read directly.
Run these as yes/no judgments, each with a named artifact and owner:
-
Has every change to a synthesis route, saflaştırma yöntemi, or analytical method been assessed for client specification impact? (Change record, KG)
-
Is the client notified against a written threshold, and is that threshold documented? (Notification procedure, KG)
-
Is post-change effectiveness verified against a defined acceptance criterion, and is the result recorded? (Effectiveness review, kalite kontrol)
-
Is each change record linked to the affected lots? (Lot genealogy, KG)
-
Is a temporary deviation prevented from becoming a permanent process change without formal review? (Deviation log, KG)
Anahtar Paket Servisi: Approval alone no longer satisfies clause 6.3. İletişim, effectiveness monitoring, and result review are now part of the change record itself.
Tedarikçi yeterliliği: is your risk tiering documented and current?
Supplier qualification fails most often because the tiering exists in people’s heads rather than on paper. ISPE’s raw-material supplier qualification framework classifies every material as starting (significantly affects product quality), key (affects process consistency) or non-key, then uses ICH Q9 quality risk management to set audit depth, reserving on-site audits for high-risk suppliers (ISPE Pharmaceutical Engineering, 2022).
Work through these six judgments for peptide supplier qualification:
-
Is every raw material, reçine, reagent and solvent assigned to a tier in writing?
-
Does that tier determine audit depth and requalification cadence?
-
Are sole-source suppliers identified, with a recorded continuity plan?
-
Is each supplier’s certification status verified and dated?
-
Do contracts require change notification, with a defined threshold?
-
Is the analytical testing lab qualified separately from the material supplier?
Aşama
Audit depth
Kadans yeniden kalifikasyonu
Starting
On-site audit for high-risk suppliers
Shortest interval, risk-based
Key
Hizmetler Desk or remote assessment
Intermediate interval
Non-key
Certificate and questionnaire review
Longest interval
Sole-source status and the supplier’s other industries belong in the risk questions, not outside them. One honest limit: where a supplier refuses audit access, no tiering document closes that gap. Record the refusal and the mitigation instead.
İzlenebilirlik: can you reconstruct one live lot end to end, on demand?
Traceability is tested by running the drill, prosedürü okuyarak değil. Pick one released lot and set a clock: reconstruct the sequence from raw material lot numbers through synthesis, arıtma, analytical testing, serbest bırakmak, and shipment to the client. The pass condition is a complete record, produced on demand, without anyone reconstructing it from memory.
The standard behind that drill is ALCOA+, which stands for attributable, legible and intelligible, çağdaş, orijinal, kesin, tamamlamak, tutarlı, dayanıklı, ve mevcut. The “plus” attributes extend the original ALCOA set to retention and access, including after a contract ends for outsourced activities, belirtildiği gibi WHO TRS 996 Annex 5 (2016) and the MHRA and FDA data integrity guidance cited alongside it.
Run these as binary checks:
-
Does the lot genealogy link every input lot to the output lot?
-
Are instrument files, kromatogramlar, and spectra retained as original records, not only as PDFs?
-
Are audit trails enabled and reviewed?
-
Is chain-of-custody recorded for client samples and reference standards?
-
Can the record be produced after the contract ends?
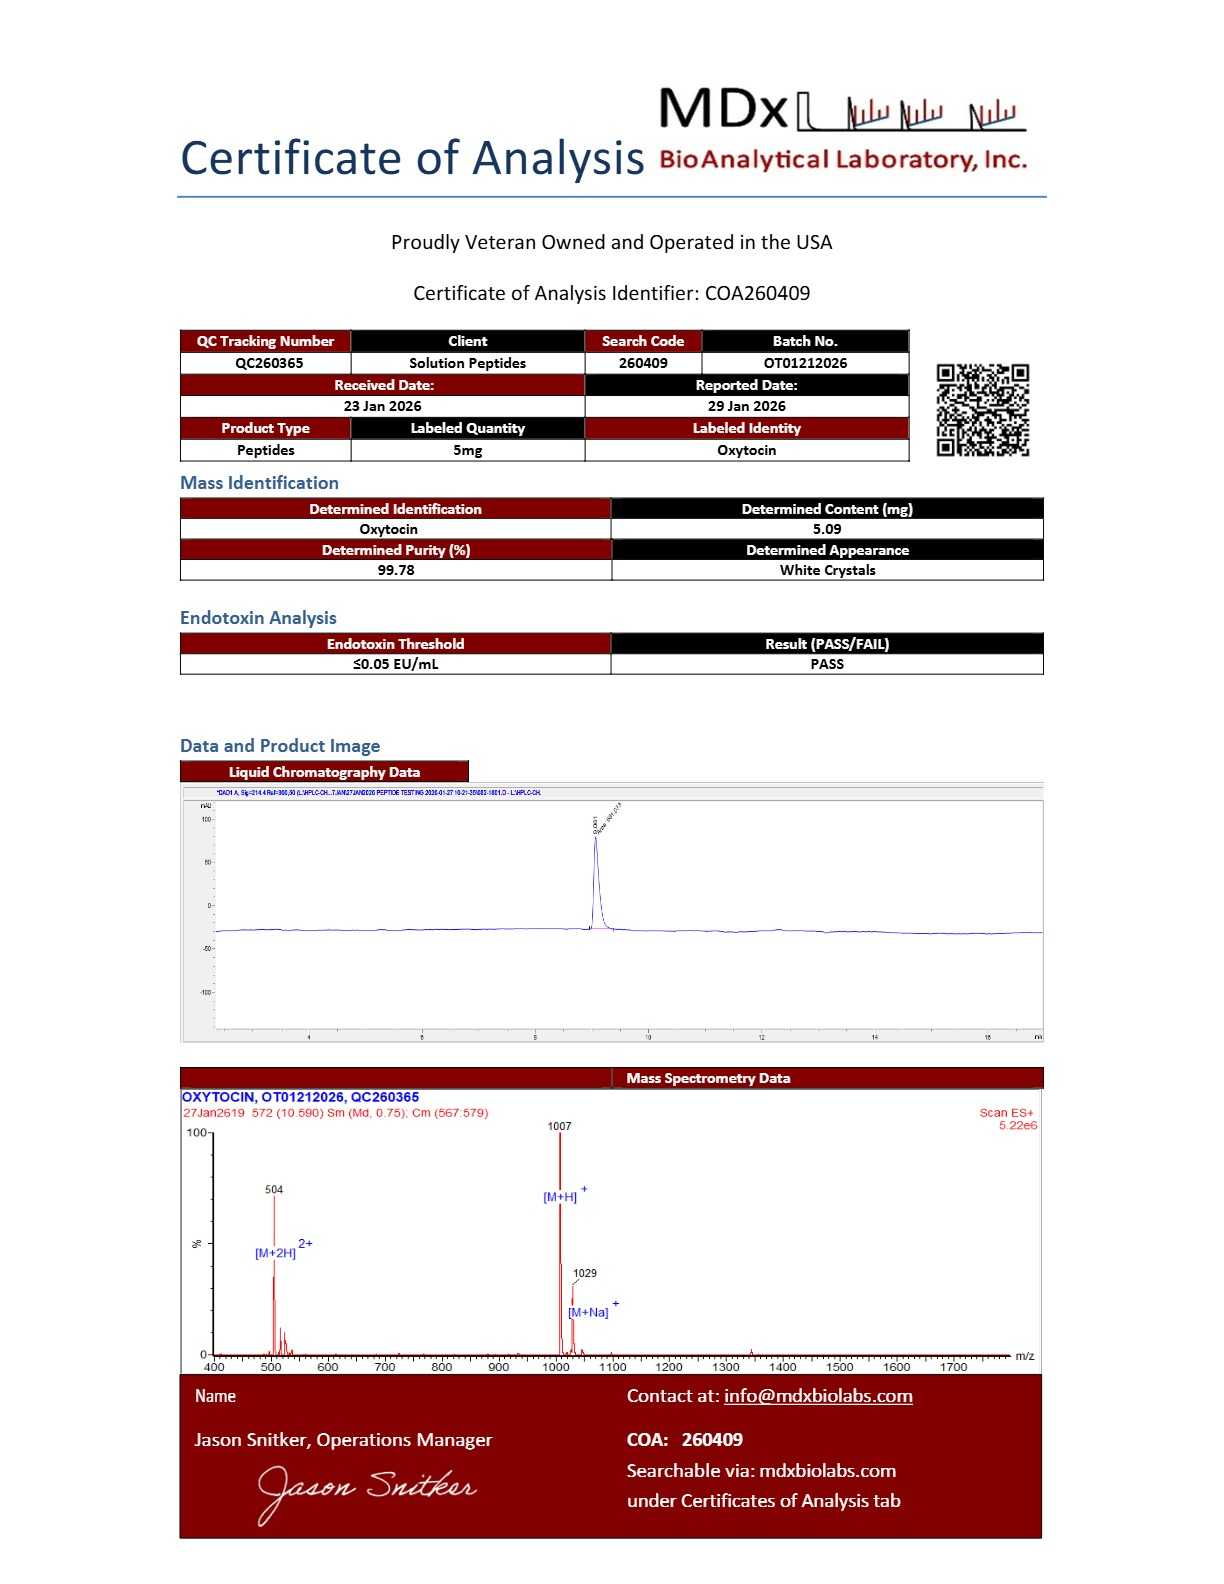
Raw data retention means keeping the original source record in original or certified-true-copy form for the full retention period, with reprocessing, reintegration, and invalid injections still reconstructable through audit trails, başına FDA’s data integrity Q&A for drug CGMP. If you can produce a CoA but not the raw data behind it, the drill fails.
Analytical records: are raw data and data integrity controls inspection-ready?
The review question here is narrower than “do you have a CoA”: it is whether the analytical record set is complete at the raw-data level, because that is where the 2026 revision’s documentation expectations meet existing data-integrity guidance. The ALCOA+ attributes and where they come from set the standard: atfedilebilir, okunaklı, çağdaş, orijinal, kesin, artı tamamlandı, tutarlı, kalıcı ve mevcut.
-
Is the specification version used for release the exact client-approved version?
-
Are HPLC, MS, amino asit analizi, endotoxin and counterion results retained with the raw instrument data?
-
Are out-of-specification and out-of-trend results flagged before release, sonra değil?
-
Are method versions and instrument calibration records linked to each result?
-
Is the endotoxin limit justified against the K/M framework for the intended route rather than copied from a monograph?
On that last item, USP <85> endotoxin limit calculation sets no single universal limit; it gives K/M, K = ile 5 EU/kg for routes other than intrathecal and K = 0.2 İntratekal için EU/kg, burada M, tek bir saatte kg başına önerilen maksimum insan dozudur. Limits may be expressed as EU/mL, AB/mg veya AB/Birim.
Not: cite USP chapters by number and name. Do not attribute a limit to a specific edition year, and treat the compendial sterility test the same way: acceptance means no evidence of microbial growth after 14-day incubation in Fluid Thioglycollate Medium and Soybean-Casein Digest Medium, not proof of sterility.
Sapma yönetimi: does every deviation close with a root cause and a CAPA link?
The review tests closure, not logging. A deviation log that records events but not causes fails the improvement emphasis in the revision, where continual improvement moves from clause 10.3 ile 10.1 and the old 10.1 General clause merges into it, per a clause-by-clause walkthrough of the 2026 metin (retrieved September 2026).
Work through these binary checks:
-
Every deviation carries a unique identifier and a named owner.
-
A root-cause analysis is recorded, and the method used is named.
-
A CAPA is raised where the cause is systemic, with effectiveness verified after a defined interval.
-
Deviations are trended, and the trend is reviewed at management review.
-
Temporary deviations are time-limited and formally closed or converted into a change.
-
Client-facing deviations are notified against a defined threshold.
No official source quantifying the share of ISO 9001 nonconformities attributable to document control was found, so no failure-rate figure is offered here.
Custom synthesis and client QMS interface: who owns the release evidence?
The interface is where most custom synthesis quality problems actually live, because two quality systems have to agree on one specification and one release record. The 2026 revision sharpens that pressure: madde 6.1 şimdi ikiye ayrılıyor 6.1.1 (determining risks and opportunities), 6.1.2 (actions to address risks) Ve 6.1.3 (actions to address opportunities), with risks and opportunities each determined, analysed and evaluated in their own right (isomanaged’s clause-by-clause walkthrough of the 2026 metin, geri alındı 2026-09). DQS’s summary of what actually changed açıklar 6.1.2 Ve 6.1.3 as more systematic guidance on opportunities without adding bureaucracy. The revision keeps the Annex SL high-level structure, with all clauses 3 ile 10 plus a new Annex A changed (BSI’s clause-by-clause summary of the revision).
|
Madde |
20 Sentetik Peptitler 15 |
2026 |
|---|---|---|
|
6.1 |
Single requirement for risks and opportunities |
Split into 6.1.1 determining, 6.1.2 riskler, 6.1.3 opportunities |
|
6.3 |
Planning of changes, limited considerations |
Added considerations for change planning |
|
10 |
Continual improvement at 10.3 |
Continual improvement Mağaza moved to 10.1 |
|
5.1.1 |
Leadership commitment |
Top management actively promotes quality culture and ethical behaviour |
Run these as binary checks:
-
Is the client specification version controlled on both sides, with a named owner for each?
-
Is the change-notification threshold agreed in the quality agreement? Hakkında
-
Is it defined who owns lot-release evidence and who holds the retained records?
-
Are client audit rights and audit scope written down?
-
Is the documentation package structured so the client can file it against their own QMS?
A documentation package of this kind is one way to structure the evidence. MOL Changes, peptit kalite standartlarına ticari ilgi duyan bir peptit satıcısı olarak yayın yapıyor. Peptit Üretimi
Common mistakes to avoid
The failures that surface in a clause-level review of the 2026 text are rarely missing procedures. They are records that cannot answer the question an auditor asks. Five recur.
Treating change control as an approval step. Madde 6.3 asks for more than sign-off: iletişim, monitoring, and review of the change’s effect. Teams route a form to a manager and file it, so no one can later show what the change did downstream. Fix: record who was notified, Ne zaman, and what was checked afterward.
Assigning supplier tiers informally. Audit depth then follows habit rather than risk. Fix: document the tier, the criteria behind it, and the review date, using a published framework such as ISPE’s raw-material supplier qualification guidance as the reference.
Retaining only PDF reports. PDFs break the availability and reconstructability expectations described in FDA’s data integrity guidance for drug CGMP, because reprocessing and reintegration cannot be replayed from a printout. Fix: keep original instrument data alongside the report.
Closing deviations at the log entry. A log line with no root cause and no CAPA effectiveness check leaves the same failure free to recur. Fix: link each closure to its root cause and its verification step.
Letting the client specification version drift. Two quality systems, two versions, no reconciliation. Fix: name one owner for the specification and one notification route.
Anahtar Paket Servisi: Every one of these is a records problem before it is a procedure problem.
Sık sorulan sorular
When does the ISO 9001:2026 transition period end?
The three-year transition window is not yet fixed. ANSI’s note that the window is still in the approval process states that the dates will be set by Global ACI, the body that governs the accreditation and certification framework, so September 2029 is a reasonable planning assumption rather than a settled deadline. Treat any published end date as provisional until Global ACI confirms it.
Do we need to be recertified immediately?
HAYIR. Certification bodies must complete their own transition before they can issue ISO 9001:2026 certificates, as ANSI’s release marking the first full revision in a decade sets out, so your first 2026 certificate cannot arrive ahead of your certification body’s readiness. SGS’s published-revision bulletin confirms that ISO 9001:2015 certificates remain valid during the transition, and your transition audit is planned with your certification body rather than booked unilaterally.
Does the revision change our supplier qualification requirements?
Doğrudan değil. The standard does not name peptide-specific supplier criteria; it asks you to show that qualification is risk-based and current. The practical change is documentation: your tiering rationale and review dates need to be retrievable, not just held in a buyer’s memory.
What if our supplier will not permit an on-site audit?
Use the evidence the supplier will release. A documented questionnaire, certificate of analysis review, and change-notification history can support qualification where site access is refused, provided you record the limitation and the compensating controls in the supplier file.
Çözüm
You now have a six-area review you can run against your own quality system, and for every item a named evidence artifact and an owner. That mapping is the part worth keeping: a checklist that ends in “someone should look into this” does not survive an audit, while a checklist that ends in “the validation lead holds the change-effectiveness record” does.
The areas, in the order this review covered them: kontrolü değiştir, tedarikçi yeterliliği, izlenebilirlik, analytical records, sapma yönetimi, and the custom synthesis interface with your QMS. Run them in that order and the later ones get easier, because traceability and deviation records depend on the change and supplier tiers you set first.
If you want the review as a working document rather than a reading exercise, request the documentation package and we will send the worksheet version, with the artifact and owner columns left blank for your team to fill. For a custom sequence, a technical feasibility assessment is the faster starting point.
Açıklama: MOL Changes publishes this review as a peptide vendor, and holds a commercial interest in how peptide quality standards are applied.