What makes peptide sourcing a scientific risk decision rather than a procurement choice?
Synthesis Of Polypeptide Factory Hgh Peptides The decision is scientific because the failure modes are analytical, and the documented consequences come from material that was never verified. Belgian regulators screened the ten most frequently encountered falsified peptide drugs on the Belgian market, obtained from three suspected illegal internet pharmacies, and found measured purity ranging between 5% 그리고 75% for cysteine-containing peptides, alongside high variation in the amount of drug per unit (Janvier et al., Talanta, 2018). The same screening found class-one elemental impurities: one sample contaminated with lead, and multiple samples with arsenic at concentrations up to ten times the ICH toxicity limit for parenteral drugs, with speciation confirming all arsenic was present in the more toxic inorganic form (Janvier et al., Talanta, 2018). Those results describe falsified products from illegal pharmacies, not legitimate API manufacturing, but they establish what an unverified source can deliver.
The criteria are also moving. FDA withdrew its May 2021 guidance on highly purified synthetic peptide drug products on 28 칠월 2026 because it no longer reflects FDA’s current scientific thinking, and published 17 revised draft product-specific guidances covering production route, 선천면역반응검사, 불순물 임계값, 고차원 구조 평가 및 생물학적 활동 평가 (FDA CDER, 28 칠월 2026). A supplier qualification built on a 2021-vintage reading of FDA’s position is therefore already out of date, which is why the checklist below is anchored to the record rather than to a threshold.
How do I confirm peptide identity, and why isn’t HPLC enough?
Identity confirmation requires a method that can distinguish the authentic analyte from non-analyte samples, which is the performance characteristic ICH Q2(R2) assigns to the identity purpose rather than to assay or purity (나는 Q2(R2), 채택됨 1 십일월 2023, with the published 30 십일월 2023 correction). The same guideline permits an orthogonal route: a lack of specificity in one procedure should be compensated by one or more other supporting analytical procedures unless appropriately justified (나는 Q2(R2), 2023). That clause is the reason a single chromatogram cannot close the identity question on its own.
Retention time and intact mass support different aspects of identity. Set mass tolerance from the instrument, 구경 측정, charge state and analytical purpose; ±1 Da is not a universal peptide identity criterion. Isomeric, positional or stereochemical questions may require additional methods.
What does a purity percentage actually mean, and when are two purity numbers comparable?
A purity number is only interpretable inside a specification, which ICH Q6B defines as a list of tests, 분석 절차에 대한 언급, and appropriate acceptance criteria that are numerical limits, 범위, 또는 설명된 테스트에 대한 다른 기준 (나는 Q6B, 단계 4, 10 3월 1999). Because every acceptance criterion is tied to a listed procedure, “98% pure” without the procedure is an unanchored claim rather than a specification.
The deeper problem is that two purity numbers can be measured on different bases. Reversed-phase HPLC purity is the fraction of detected UV signal assigned to the target peak under one method, while mass balance, or net peptide content, is the mass fraction of actual peptide in the weighed material and must account for water, counterions such as TFA or acetate, residual solvents and other non-peptide material; the two can diverge by several percent (McCarthy et al., Pharmaceutical Research, 2023). The same work assigns purity by subtracting all detectable impurities, 반대이온, 물, residual solvents and non-combustible residues from 100%, and reports the least inter-laboratory variability when the mass-balance approach is used (McCarthy et al., 2023). The thresholds that circulate as if they were requirements, 0.1% for identifying an individual impurity and 0.5% for total impurities, are attributed to FDA drafted guidance for synthetic peptides (워터스 애플리케이션 노트 720006367, 팔월 2018), while FDA’s peptide team states that no official qualification levels exist (FDA OGD/CDER, 2013). Treat them as draft-stage practice, not as a pass mark.
A purity claim is comparable only when it carries five fields: the method and column, 그라데이션, the detection wavelength, the integration rules, and the reference standard used. Ask for all five before comparing two suppliers’ numbers.
What must a batch-specific COA contain, and how do its records connect?
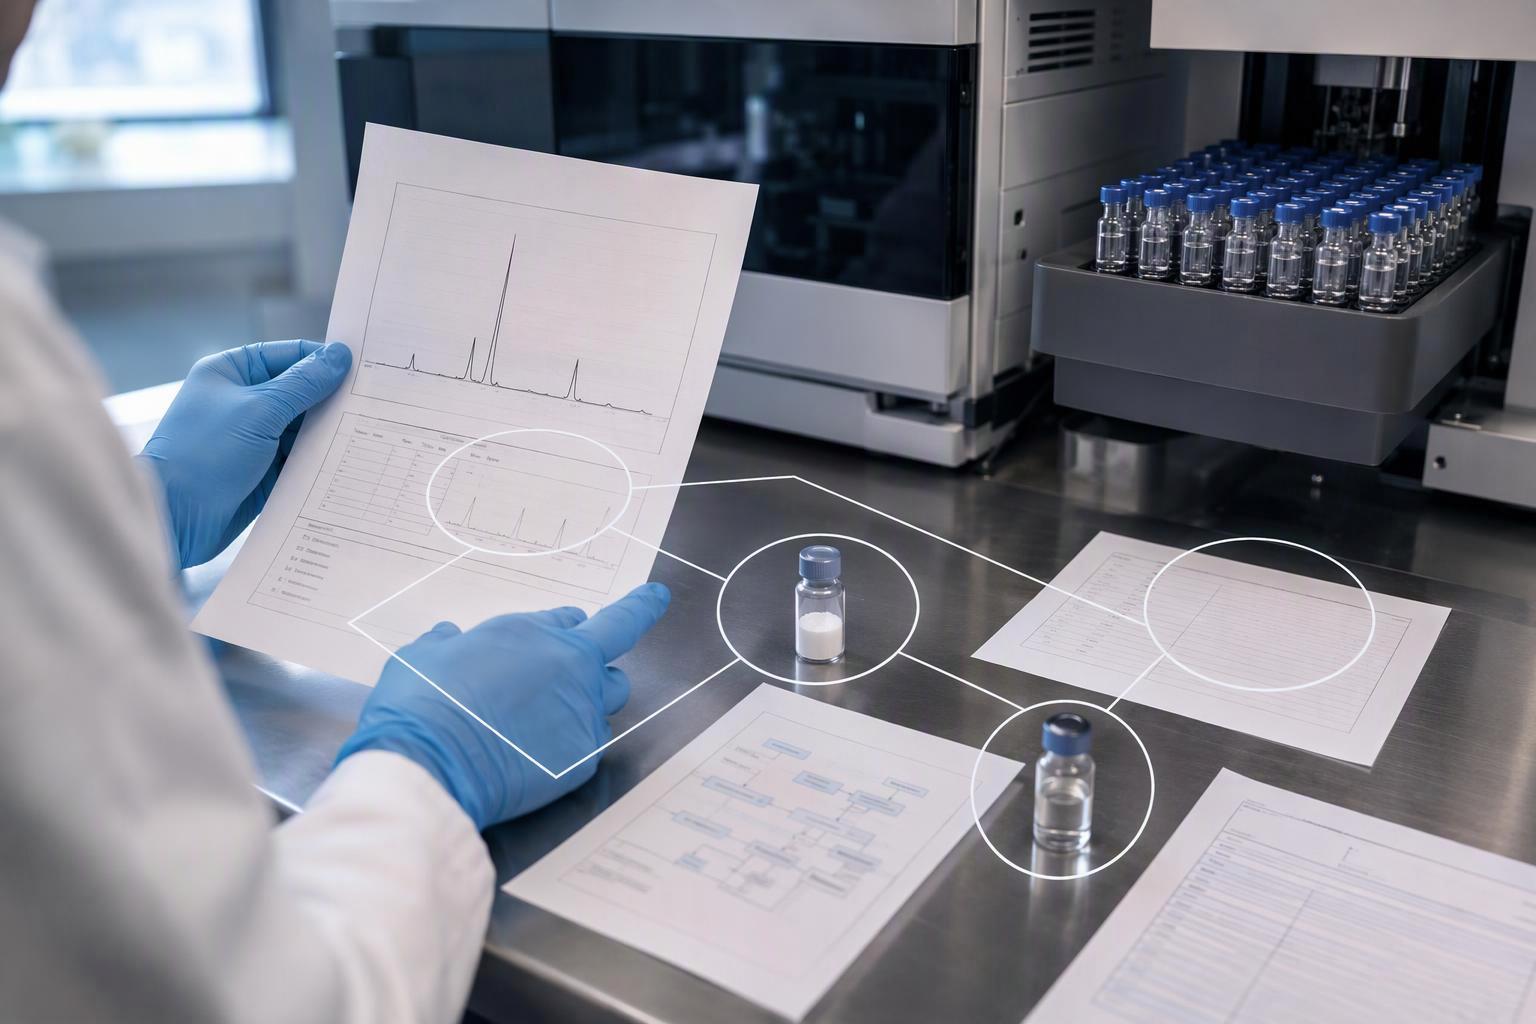
A batch-specific CoA summarizes specified tests and results. Supporting chromatograms, 스펙트럼, methods and source records should Peptide S Supplier be available at the depth required by the use and quality agreement; a CoA need not embed every raw analytical file.
The records then have to connect to one another. Lot traceability links the product label, the COA, the receiving record and the inventory record, and a strong supplier signal is a lot number that matches across all four, against a weak signal of no lot-level documentation trail at all (kb://41779). Identity testing is documented when HPLC, LC-MS, mass spectrometry or a related analytical method is listed, and absent when no analytical method is stated (kb://41779). The data behind those records is governed by 21 CFR 부분 11, which requires secure, 컴퓨터 생성, time-stamped audit trails that independently record the date and time of operator entries and actions creating, 전자 기록 수정 또는 삭제, with changes that do not obscure previously recorded information (21 CFR §11.10(이자형), 최신판). The same part defines an electronic record as any combination of text, 제도법, 데이터, 오디오, pictorial or other information representation in digital form created, 수정됨, 유지되다, 보관된, retrieved or distributed by a computer system (21 CFR §11.3(비)(6), 최신판). A COA that arrives as a scanned image with no retrievable source record fails that standard.
How do I judge lot-to-lot consistency before I commit to a supplier?
Assess consistency with comparable measurements across relevant lots, 검증되거나 검증된 방법, reference materials and change history. General ICH registration batch counts and stability durations are not mandatory evidence for every research-peptide purchase; select a study design appropriate to the intended use.
The reference-standard literature shows why the batch question is not academic. In USP’s work, gonadorelin was highly stable under thermal stress while bivalirudin was more susceptible and showed significant degradation at higher temperatures (McCarthy et al., 2023). Two peptides handled identically behaved differently, so a stability statement transferred from one peptide to another carries no weight. For scale-up specifically, the documented practice for projects in the 1–100 g range is that HPLC purification is highly recommended, endotoxin testing is standard, and comprehensive batch records are maintained for traceability, with a per-batch panel covering LC-MS, UPLC duplex analysis, endotoxin assay, sodium content, 수분 함량, 바이오버든, UV assay, residual solvent analysis and sterile filtering (kb://28940). Ask which of those tests are performed per batch and which are performed once.
How do storage, shipping and handling enter the quality record?
Storage and shipping enter the quality record because the conditions a supplier claims must be justified by that substance’s own stability data rather than by a universal rule. 나는 Q1A(R2) sets the study conditions and the minimum data package, and treats drug substances intended for storage below −20 °C on a case-by-case basis (나는 Q1A(R2), 2003). Data from the accelerated condition can be used to evaluate the effect of short-term excursions outside the label storage conditions, such as might occur during shipping (나는 Q1A(R2), 2003). That is the clause that makes a shipping qualification a document rather than a promise.
Supplier handling guidance fills in the practical detail. Lyophilised peptide should be stored tightly closed below −15 °C, with lower temperatures such as −50 °C or below for long-term storage; the container should reach ambient temperature in a desiccator before opening because peptides are hygroscopic and moisture uptake reduces both peptide content and stability; and material should be weighed out quickly and the container tightly resealed (Bachem handling and storage guidelines, supplier technical documentation). The same guidance notes that peptides containing Asn, Gln, 만난, Cys or Trp have limited shelf lives, and states that no standard protocol for dissolving a peptide can be provided because properties vary extraordinarily (바헴). A checklist item here is binary: does the supplier state the storage condition, the product form and the handling instruction for the specific lot, or only a generic statement.
What disqualifies a peptide source as unverifiable?
If the supplier cannot connect a received lot with relevant identity, purity and content results, record the evidence gap and defer acceptance where those Polypeptide results are required. This is a proposed procurement criterion, not an external standard or a conclusion based on an inaccessible internal reference.
The “research use only” label is not a substitute for that record. FDA의 26 2월 2025 warning letter to USApeptide.com found unapproved and misbranded semaglutide and tirzepatide introduced into interstate commerce despite labelling stating “research use only,” “not for human consumption” and “lab purposes only,” because the site’s own content established intended human use (FDA warning letter MARCS-CMS 696885, 26 2월 2025). The same letter states that unapproved drugs may be contaminated, counterfeit, contain varying amounts of active ingredients, or contain different ingredients altogether (FDA, 2025). The disqualifying rule is therefore: if the supplier cannot supply a batch-specific COA with the chromatogram and the mass spectrum, a lot number that matches the label and the receiving record, and the storage and handling documentation for that lot, do not order. Request the batch documentation package and put the three documents side by side before committing.
자주 묻는 질문
Does a research-use-only label change what a supplier has to document?
아니요. RUO positioning describes the intended research context, not a reduced paperwork standard. The procurement review still turns on the same records: a batch-specific certificate of analysis, documented identity and purity information, a traceable lot number, a clearly stated product form, and storage and handling guidance available before intake. A supplier that treats the RUO label as a reason to withhold batch-level records is offering less, not something different. 맞춤형 펩타이드 합성 도매
Can our own HPLC run replace the supplier’s identity data?
It supplements it, but it does not substitute. An in-house run confirms what your method was built to detect, on your column and your reference standard, and it tells you nothing about how the material was characterized at release. Keep the supplier’s identity record as the release documentation and treat your own analysis as an independent check against it. Where the two disagree, the discrepancy itself is the finding worth chasing. Planet Peptide
What if the supplier offers a generic COA but not a batch-specific one?
Treat that as a gap you cannot close from your side. A generic certificate describes a product line; a batch-specific certificate of analysis ties the identity, purity and method records to the lot in the vial you received. Without the lot-level link, you cannot reconstruct what was measured, 언제, or against which reference standard. Ask for the batch document, and if it is not available, record the absence rather than accepting the generic version as equivalent.
Do catalog peptides need the same documentation depth as custom sequences?
예, with one difference in emphasis. Catalog items are usually characterized once per lot against a fixed specification, so the batch-specific COA and lot traceability carry most of the weight. Custom sequences add synthesis-specific variables, including modification pattern, terminal groups and purification route, so the documentation has to describe the actual delivered molecule rather than a catalog entry. Both cases still require identity confirmation and purity records at the batch level.
How should counterion form be specified?
Write it into the contract, 자격증 뿐만 아니라. Trifluoroacetate is the default salt for many syntheses, and assuming acetate when the material is supplied as the TFA salt can invalidate TFA-sensitive cell-based assays, which is why counterion form belongs in the contract as well as the COA. State the required salt form, require it to appear on the batch document, and treat a mismatch as a specification failure rather than a labelling detail.
Are sterility and endotoxin results interchangeable?
아니요. Sterility and endotoxin are separate tests, and a peptide can pass one while failing the other, because killing organisms does not remove pyrogenic lipopolysaccharide (USP <85> adoption notice, official date 1 12월 2012). If your intake criteria call for both, request both as distinct line items on the batch record.
결론
The checklist reduces to five document requests, one per category, and a single disqualifier. 정체성을 위해, ask for the orthogonal method set rather than a single peak. For purity, ask for the five comparability fields and treat the impurity figures as draft-stage practice, since FDA’s peptide team states no official qualification levels exist for peptide-related impurities (FDA OGD/CDER, 2013). For content, ask for net peptide content alongside the HPLC area percentage, because the two are measured on different bases (McCarthy et al., 2023). For stability, ask for multi-batch data at the conditions ICH Q1A(R2) 세트, including the shipping excursion assessment (나는 Q1A(R2), 2003). For assay suitability, ask which tests are run per batch and which are run once. If the batch-specific COA, the lot number trail and the storage documentation for that lot cannot be produced together, the source is unverifiable and the order stops there.