Selección de péptido CRO para fase 1: Los criterios que sobreviven a una IND acelerada
Selección de péptido CRO para la fase 1 ahora ocurre en un contexto regulatorio en movimiento, y los criterios que importan no se han movido con él. El HHS lanzó la Operación TrialBlazer en junio 22, 2026 como un esfuerzo de todo el departamento que abarca la FDA, NIH, ARPA-H, OIG y ONC (comunicado de prensa del HHS, 2026-06-22). Debajo de él, Programa piloto IND acelerado de la FDA permite a los patrocinadores asociarse con instituciones de investigación calificadas para que la FDA pueda aceptar y revisar componentes IND individuales de forma continua a medida que se completan..
Lo que cambia el piloto es el mecanismo de revisión. Lo que deja intacto es lo que la FDA espera en una fase inicial 1 INDIANA: información suficiente para asegurar la correcta identificación, calidad, pureza, y potencia del fármaco en investigación, con la cantidad variando según la fase, formulación, y duración.
Califique a cada CRO candidato según si puede generar esa evidencia, no sobre si puede mover un envío más rápido. Como "Más allá del reloj IND" del líder clínico argumenta, el reloj de revisión del IND rara vez es la limitación vinculante. El cuello de botella se sitúa en todo lo que transcurre entre la preparación del IND y la administración de la dosis al primer paciente., donde el piloto no llega.
Conclusión clave: El piloto comprime la logística de revisión, no expectativas de CMC. Un CRO que no puede defender la identidad, pureza, y los datos de fuerza no serán rescatados mediante una presentación continua.
Qué cambia realmente la operación TrialBlazer para los programas de péptidos
El elemento realmente nuevo es la secuenciación de revisiones., no es un estándar de evidencia más bajo. bajo el piloto, Los patrocinadores pueden solicitar unirse hasta 11:59pm ET en octubre 30, 2026, y la FDA revisa los componentes IND completados de forma continua en lugar de esperar la presentación completa (Página de acciones de la FDA, recuperado 2026-09-15). El razonamiento declarado por la FDA es que la revisión continua le brinda “una oportunidad más temprana de resolver posibles deficiencias que darían lugar a una suspensión clínica o una solicitud de información”.,” para que la carta de seguridad para proceder pueda emitirse antes (arnold & Aviso de portero, 2026-06-24).
Dos límites importan cuando lees las afirmaciones de los proveedores. Las instituciones de investigación calificadas actúan únicamente como “recursos de revisión y asesoramiento”: los patrocinadores “seguirían siendo responsables de sus presentaciones IND," y la FDA "mantendría la supervisión regulatoria total de la presentación del IND, incluida la autoridad para imponer una suspensión clínica” (arnold & Aviso de portero, 2026-06-24). Ausencia de una carta de seguridad para continuar o de una retención, el plazo legal de 30 días a partir de la recepción del IND todavía rige. La mecánica y los límites propuestos se abrieron para comentarios públicos en 91 alimentado. registro. 37996, así que trátelos como una propuesta y no como un procedimiento establecido..
Los cuatro fundamentos La velocidad no se relaja
La operación TrialBlazer comprime la línea de tiempo de IND, no las expectativas de CMC. Solicitudes IND de la FDA: La página de información de CMC enmarca la presentación en torno a cuatro fundamentos que se aplican en cada fase.: identificación, calidad, pureza, y fuerza. La cantidad de información que respalda cada uno varía según la fase., la formulación, y la duración prevista del estudio., pero ninguno de los cuatro desaparece porque se agiliza un programa.
Lo que la FDA relaja es más limitado de lo que parece en un principio. La agencia enumera cuatro fases explícitas 1 flexibilidades: Es posible que no se requieran datos de estabilidad del lote clínico específico., Es posible que no sea necesario que los datos de estabilidad iniciales cubran la duración total de los estudios propuestos., No se esperan datos de validación del método analítico., y no se espera que se especifiquen controles de proceso que no estén directamente relacionados con la seguridad del producto..
|
Fase 1 Artículo CMC |
Debe enviar |
puede diferir |
|---|---|---|
|
Identidad, calidad, pureza, fortaleza |
Sí, a la profundidad apropiada para la fase |
Profundidad de caracterización, no la caracterización en sí |
|
Estabilidad específica del lote |
No requerido para el lote clínico. |
Respaldado por plataforma o datos de lote representativos |
|
Duración de la estabilidad |
No es necesario cubrir el estudio completo. |
Aún debe soportar el período de espera clínica. |
|
Validación del método analítico |
Servicios No esperado |
Aún se espera la calificación apropiada para la fase |
|
Controles de proceso no relacionados con la seguridad. |
No requerido |
Los controles vinculados a la seguridad siguen vigentes |
“Not expected” is not “prohibited.” A CRO that reads the flexibility as license to skip characterization is the failure mode this framework exists to catch, because the same four fundamentals return in force at Phase 2.
How to Evaluate a Peptide CRO’s Identity and Purity Capability
The first question to settle in a proposal review is not what platform a CRO runs, but whether its release method can resolve the impurities your sequence actually produces. FDA’s expectation for an initial Phase 1 IND is that the sponsor identifies the drug substance and controls its purity, which is what FDA expects in an initial Phase 1 INDIANA. Peptide identity and purity testing therefore has to stand on more than one leg.
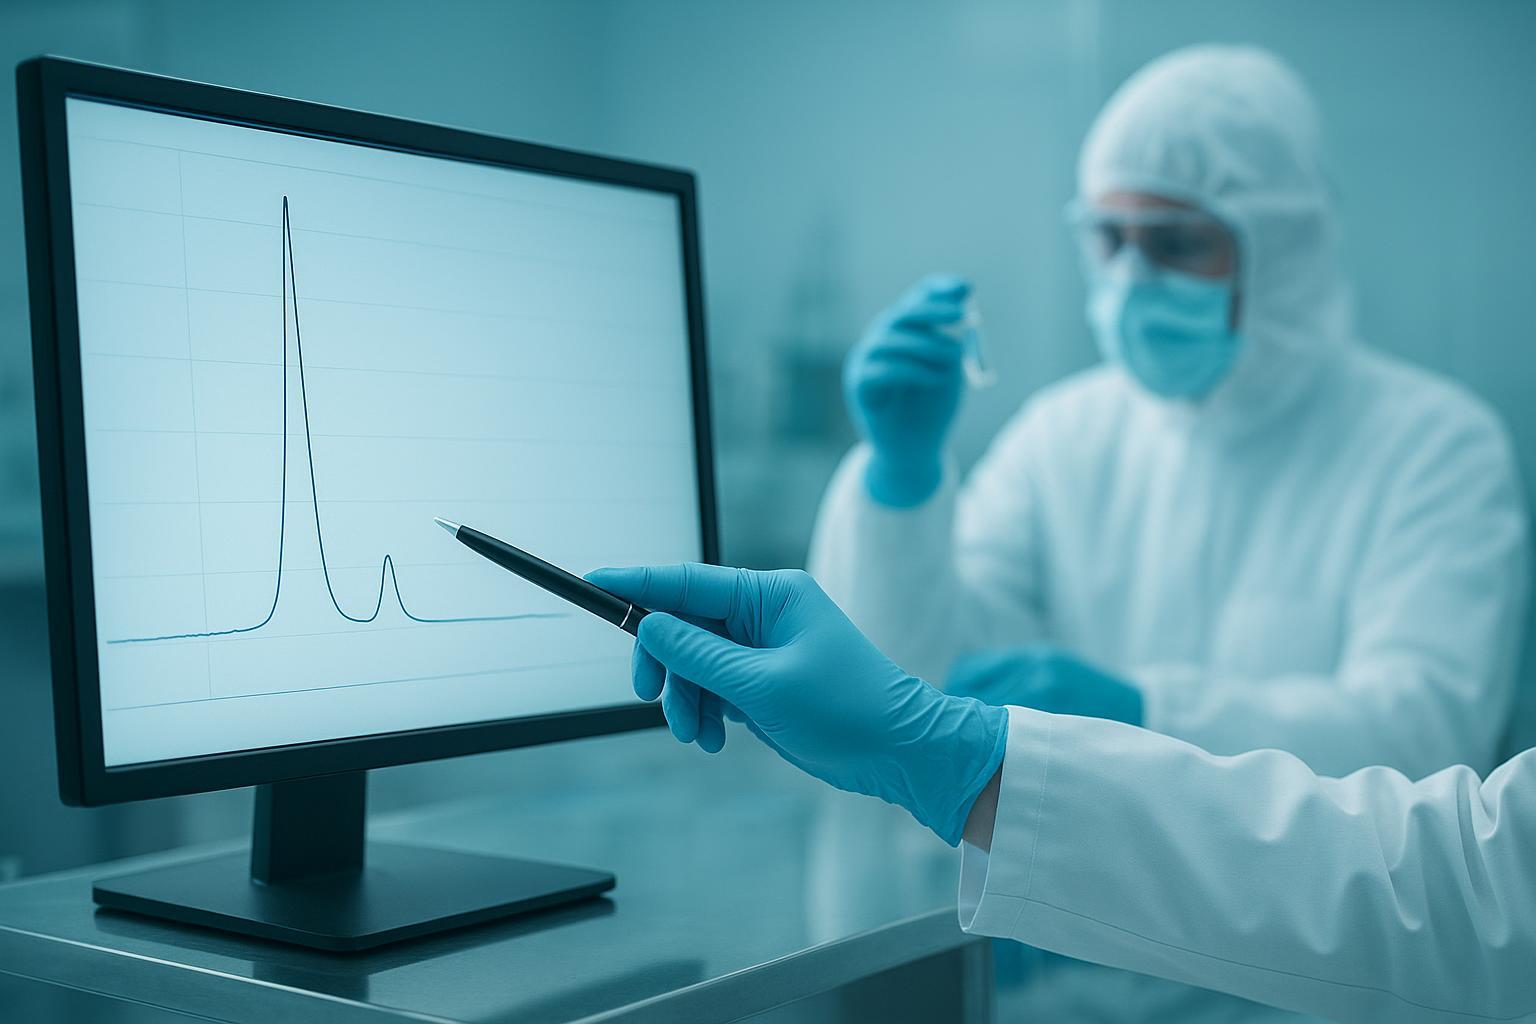
Ask for orthogonal confirmation: mass spectrometry for molecular mass, plus chromatographic relative retention against a reference standard. For sequences where truncations, isómeros, or deamidation products elute near the parent, request peptide mapping or amino acid analysis as the second identity method, and ask to see a representative chromatogram with the impurity peaks labeled.
The failure mode is specific. A CRO that offers a single identity method, or that cannot resolve a 40-residue hydrophobic truncation impurity from the parent peak on the release method, will leave you defending an unresolved peak in the IND.
Conclusión clave: Two orthogonal identity methods and a demonstrated truncation separation are the minimum evidence that a peptide CRO’s release testing will hold up in a Phase 1 INDIANA.
Cómo evaluar métodos indicadores de estabilidad y validación apropiada para la fase
Full analytical method validation data are not expected at Phase 1, so the question is not whether you can defer validation work. It is which deferred data still has to be defensible later.
The distinction is stability-indicating capability. A method can be unvalidated and still be stability-indicating, but only if it demonstrably detects degradation, and the usual proof is forced degradation: stress the peptide under acid, base, oxidación, heat and light, then show the method resolves the degradants from the parent. Ask a candidate CRO for that evidence before you ask for a validation package. A CRO that cannot produce forced-degradation data is offering you a method whose ability to see degradation is simply unknown, and that gap surfaces when a later phase asks you to defend the assay you have been releasing against.
FDA has been explicit about what FDA says sponsors over-submit: six months or more of stability data when only trial-duration coverage is needed, and it states that “a fully developed and validated commercial process is unnecessary at this stage.” The same source notes FDA intends to let sponsors reuse manufacturing and testing data from one product to support the next where methods are standardized and well understood. That is the practical test for phase-appropriate method validation for peptides: standardized, transferable methods earn reuse, bespoke ones do not.
Where the pilot does not reach is worth stating plainly. The Clinical Leader analysis of Operation TrialBlazer reports that pre-IND meeting request to IND submission in the US averages 380 días, with a range approaching 700, and that site activation routinely exceeds 160 days against the National Cancer Institute’s 90-day gold standard. Those intervals sit outside the review clock the pilot compresses, and no analytical strategy shortens them.
Our finding: For peptide stability-indicating methods, the deferrable item is the validation package. The non-deferrable item is evidence that the method detects degradation at all.
Cómo evaluar la endotoxina, Esterilidad, y especificaciones de liberación de péptidos
Dose and route set the endotoxin specification, not a house default. The FDA’s inspection technical guide on Bacterial Endotoxins/Pyrogens defines the limit as K/M, where K is 5.0 EU/kilogram for parenteral products and 0.2 EU/kilogram for intrathecal routes. The same guide works the arithmetic for a 1000 mcg/ml product at a 14.3 mcg/kg dose: 5.0 ÷ 14.3 = 0.35 EU/mcg, cual es 350 EU/ml.
That number is a property of your protocol, so a CRO quoting a fixed limit has not read your dose. Ask for the calculation against your actual clinical dose and route, with the compendial methods named: USP <85> for bacterial endotoxins, USP <71> for sterility, both inside the 21 CFR 211 GMP frame.
Síntesis de péptidos Water controls deserve the same scrutiny. The FDA guide sets Sterile Water for Injection at 0.25 EU/ml and Bacteriostatic Water for Injection at 0.5 EU/ml, so a diluent switch mid-program can move your release specification without anyone flagging it.
El modo de falla es silencioso.: a default limit applied to a peptide that later moves to an intrathecal or higher-dose presentation, discovered at release testing when the lot is already made.
Construyendo la matriz de decisiones: Puntuación de CRO de péptidos candidatos
Selección de péptido CRO para la fase 1 works best when every proposal is scored against the same criteria rather than read on its own terms. Build the matrix before you open the first proposal.
|
Criterion |
Pass condition |
Modo de falla |
|---|---|---|
|
Identity and purity testing |
Orthogonal methods with system suitability data |
A single method carrying the Comercio whole release decision |
|
Métodos indicadores de estabilidad. |
Forced degradation shows the method separates degradants |
Method validated only on the fresh standard |
|
Phase-appropriate validation |
Validation depth matched to Phase 1 usar |
Phase 2-grade validation sold into a Phase 1 timeline |
|
Endotoxin and sterility |
Limits and methods stated per route of administration |
Limits copied from an unrelated product |
|
Data package readiness |
Raw data, metodos, and reports transferable as-is |
Data locked in a proprietary format |
Score each candidate pass or fail per row, then weight the rows by how much rework a failure would cost you. A vendor that fails identity and purity testing costs you a repeat study. A vendor that fails data transferability costs you the IND clock.
The two anchor texts worth reading before you score anything are what FDA expects in an initial Phase 1 IND and what FDA says sponsors over-submit. Both shape which rows belong in the matrix at all.
Errores comunes que cometen los patrocinadores en un cronograma acelerado
Treating “analytical method validation data are not expected” as permission to skip characterization. The FDA guidance for the Expedited IND Pilot states that analytical method validation data are not expected in the initial IND submission, which sponsors sometimes read as a waiver of the underlying work. No lo es. Identidad, pureza, and impurity characterization still have to exist before you can release a peptide lot for dosing, and the pilot only changes when the paperwork arrives, not whether the data exist. The fix is to keep characterization on the critical path and treat the IND as a reporting milestone rather than a laboratory one. Acerca de
⚠️ Advertencia: “Not expected in the submission” describes the filing, not the bench. A sponsor who stops characterization to save time will discover the gap at the release method, when a 40-residue hydrophobic sequence fails to resolve from its truncation impurity.
Submitting six months or more of stability data when trial-duration coverage is enough. Sponsors default to the longest stability package they have because that is what a standard IND review has historically rewarded. bajo el piloto, the relevant question is whether the data cover the duration of the proposed trial, and a package built to that scope is the one that matches the pilot’s proposed mechanics and limits. The fix is to map your stability commitment to your dosing period and hold the longer datasets for the annual report.
Assuming the qualified research institution reviews the IND. The QRI role is advisory. Sponsors who route their submission through an institution expecting a second reviewer wait for feedback that is not coming, and the wait lands squarely in the window the pilot was meant to compress. The fix is to treat QRI input as a design-stage consultation and keep your own regulatory lead accountable for submission readiness.
Assuming the pilot shortens the review clock. The binding constraint for most peptide programs sits between IND readiness and first patient dosed, where manufacturing slots, method transfer, and site activation consume the calendar. The pilot compresses the agency’s review window, a period that is often not the longest one in the sequence. The fix is to plan the pilot against your own critical path and identify where the pilot does not reach, then staff those gaps separately.
Each of these four mistakes shares a root cause: reading a flexibility as a waiver. Operation TrialBlazer Phase 1 requirements relax timing and reporting expectations in specific, named places, and every one of those places assumes the underlying science is complete.
Cómo se ve el éxito al final de la selección de CRO
Success is a scored decision matrix with a named CRO, not a shortlist. The matrix should show a documented analytical package covering identity, pureza, métodos indicadores de estabilidad, and release specifications derived from the dose and route, with each criterion scored against the same evidence you requested from every candidate.
The package should also be built for reuse. FDA has said sponsors over-submit data that is already well understood and standardized, which is the same logic behind what FDA says sponsors over-submit: when methods are standardized and phase-appropriate Péptidos sintéticos method validation for peptides is documented properly, the same package can support a later program instead of being rebuilt. That is the stretch goal worth naming now, because method-transfer readiness is where mg-to-kg scale-up drift shows up first.
Conclusión clave: You are done when you have a scored matrix, a named CRO, and an analytical package you could hand to a second program with minimal rework.
If you want to compare that package against a specific program, request the analytical and QC documentation package or speak with an analytical lead. MOL Changes provides peptide development and analytical services; this article is educational and does not recommend any specific provider.
Preguntas frecuentes
¿Aún puedo solicitar unirme al piloto IND acelerado después de octubre? 30, 2026?
The request window closes at 11:59pm ET en octubre 30, 2026, so a sponsor that has not submitted a request by then cannot join through that route. The pilot is also voluntary: it is an optional program layered on top of existing IND requirements, not a new pathway rule that governs peptide programs generally. Sponsors who miss the window still file under the standard IND framework, and the underlying Phase 1 expectations described above apply either way.
¿Trabajar con una institución de investigación calificada transfiere la responsabilidad del IND??
No. Qualified research institutions act as a review and advisory resource, not as a substitute sponsor. The sponsor remains responsible for its own IND submission, and FDA retains full oversight of the program, including the authority to place a study on clinical hold. Treat QRI input as expert review that strengthens your package, not as a transfer of regulatory accountability. Producción de péptidos
¿La fase 1 La flexibilidad en la validación de métodos analíticos se extiende a la fase. 2?
Plan for validation scope to expand rather than carry over. The flexibilities are stated for Phase 1, and the amount of information FDA expects varies with the phase of development, la formulación, and the planned duration of dosing. A method that was adequate to support an expedited first-in-human study is not automatically adequate later, so sponsors should map the validation work they will need at each stage before locking a CRO contract.
¿Cómo cambia el límite de endotoxinas para un péptido intratecal??
The calculation tightens sharply, because the K factor in the endotoxin limit calculation drops from 5.0 EU/kg for the intravenous route to 0.2 EU/kg for intrathecal administration. At the same dose, that is a 25-fold reduction in the allowable limit, which means the release specification has to be recalculated against both the route and the maximum human dose rather than carried over from an earlier program. Confirm the arithmetic with your CRO before the specification is fixed.
Conclusión
A criteria-based peptide CRO selection for Phase 1 keeps identity, pureza, estabilidad, and method-validation scope intact while the timeline compresses. The four fundamentals do not move because a program enters an expedited pathway, and the evaluation work you did against them is what protects the IND from a clinical hold later.
The upside is real but bounded. FDA’s own framing of the Phase 1 CMC flexibilities states that sponsors of first-in-human Phase 1 INDs can reduce application development time by up to 12 meses (FDA, Fase 1 IND CMC Flexibilities, recuperado 2026-06-18). That figure is an agency estimate of the ceiling, not a guarantee for any single peptide program, and it only materializes when the analytical package holds up under review.
The next step is a documentation request, not a commitment. Ask each shortlisted CRO for its phase-appropriate validation package, its impurity and truncation strategy, and a representative release specification for a peptide of comparable length and hydrophobicity, then score the responses against the matrix you built.