आवश्यक शर्तें: किसी आपूर्तिकर्ता का मूल्यांकन करने से पहले आपको क्या तैयार रखना चाहिए
A documentation walk-through takes roughly half a day per supplier. You do not need to run a full audit to get value from it, but you do need five things in front of you before the first call.
क्या तैयार रखना है:
-
आपूर्तिकर्ता का वर्तमान प्रमाणपत्र, इसकी समाप्ति तिथि के साथ
-
उस प्रमाणपत्र से जुड़ा दायरा विवरण
-
आपका गुणवत्ता समझौता, हस्ताक्षरित या ड्राफ्ट में
-
नवीनतम ऑडिट रिपोर्ट, यदि कोई मौजूद है
-
एक लाइव लॉट के लिए विश्लेषणात्मक पैकेज
Assume you are comfortable with quality-management vocabulary: अवज्ञा, सुधारात्मक कार्रवाई, प्रलेखित जानकारी. यदि वे शर्तें अभी तक दूसरी प्रकृति की नहीं हैं, the आईएसओ 9000:2015 शब्दावली साझा संदर्भ है, और कॉल से पहले इस पर सहमत होना उचित है. आईएसओ 9000:2015 गैर-अनुरूपता को "एक आवश्यकता की पूर्ति न होना" और सुधारात्मक कार्रवाई को "गैर-अनुरूपता के कारण को खत्म करने और पुनरावृत्ति को रोकने की कार्रवाई" के रूप में परिभाषित किया गया है। Most stalled supplier conversations come down to two parties using different words for the same event.
टिप के लिए: प्रमाणपत्र का दायरा विवरण मांगें, सिर्फ प्रमाणपत्र नहीं. A certificate without its scope tells you a system was certified, वह नहीं जो वह सिस्टम कवर करता है.
आईएसओ कैसा है 9001:2026 संक्रमण समयरेखा आपकी योग्यता की समय सीमा निर्धारित करती है
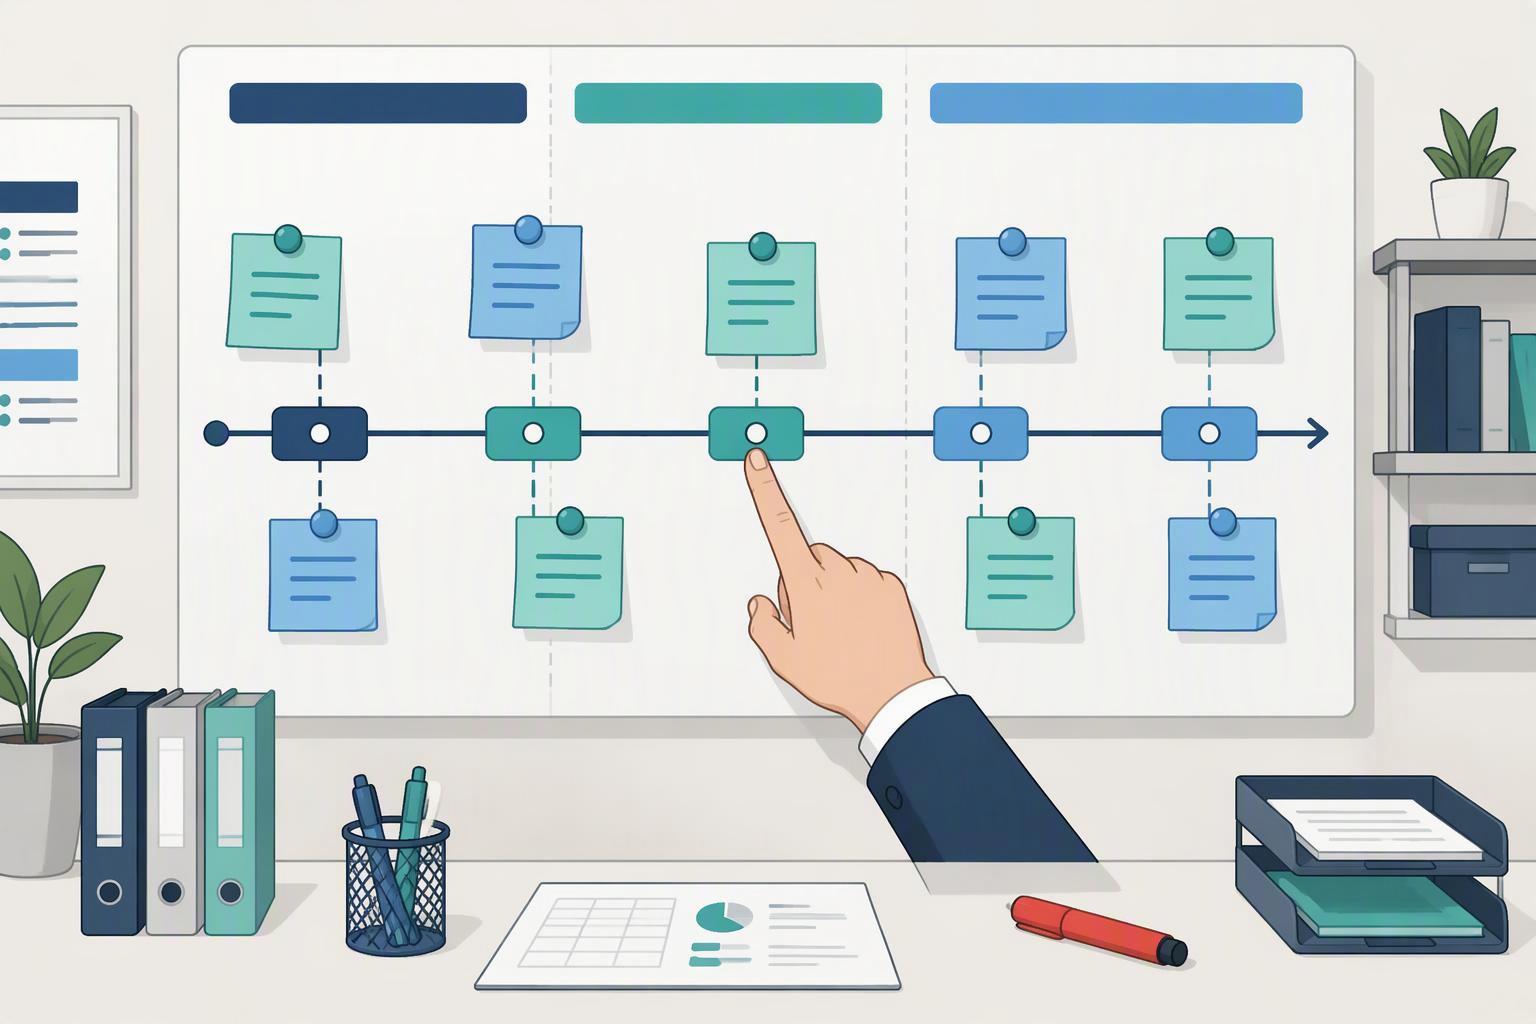
उन पाँच वस्तुओं को हाथ में लेकर, अगला प्रश्न समय का है: आईएसओ 9001:2026 transition timeline gives you four procurement deadlines before the standard’s 30 सितम्बर 2029 समाप्ति तिथि, और सबसे पहले एक गिर जाता है 31 मार्च 2027, जब मान्यता निकायों को संगठनों के विरुद्ध मूल्यांकन करने के लिए तैयार रहना चाहिए 2026 संस्करण (आईएसओ/टीसी 176 SC2 संक्रमण सूचना, पुनर्प्राप्त 2026-09-28). इसके बाद प्रमाणन निकाय अपनी संक्रमण घोषणाएँ प्रस्तुत करते हैं 30 जून 2027, मान्यता-निकाय परिवर्तन निर्णयों के साथ 30 सितम्बर 2027 (आईएसओ/टीसी 176 SC2 संक्रमण सूचना, पुनर्प्राप्त 2026-09-28).
वह तारीख जो आपके आपूर्तिकर्ता की शॉर्टलिस्ट को बदलती है 31 मार्च 2028: उस बिंदु से, नया या प्रारंभिक मान्यता प्राप्त प्रमाणीकरण केवल इसके विरुद्ध जारी किया जाता है 2026 संस्करण (आईएसओ/टीसी 176 SC2 संक्रमण सूचना, पुनर्प्राप्त 2026-09-28). आईएसओ से प्रमाणित आपूर्तिकर्ता 9001:2015 उसके बाद की तारीख अस्तित्व में नहीं रह सकती. विंडो प्रकाशन से तीन वर्ष तक चलती है, समापन 30 सितम्बर 2029 धारण करने वाले संगठनों के लिए 2015 प्रमाण पत्र, एलआरक्यूए की रिलीज़-डे ब्रीफिंग की अवधि इसकी पुष्टि करती है.
कुंजी ले जाएं: यदि आपके आपूर्तिकर्ता का प्रमाणपत्र समाप्त हो रहा है 2028 या 2029, उनका संक्रमण दायित्व पहले से ही आपके गुणवत्ता अनुबंध में लिखा होना चाहिए, और कोई भी नया आपूर्तिकर्ता जिसे आप अर्हता प्राप्त करते हैं 31 मार्च 2028 के लिए प्रमाणित पहुंच जाएगा 2026 संस्करण.
आईएसओ पुनरीक्षण को पुनर्लेखन के बजाय एक लक्षित अद्यतन के रूप में तैयार करता है, नेतृत्व पर जोर देना, गुणवत्तापूर्ण संस्कृति और नैतिक व्यवहार, जोखिमों और अवसरों पर स्पष्ट विचार, आवश्यकताओं के आशय को स्पष्ट करने वाला एक नया अनुभाग, और नवीनतम सामंजस्यपूर्ण संरचना को अपनाना (ISO का रिलीज़ नोट, 16 सितम्बर 2026). संशोधित जोर की अपेक्षा करें, प्रतिस्थापित गुणवत्ता प्रबंधन प्रणाली नहीं.
अभ्यास 1: परिवर्तन नियंत्रण जो संचार और प्रभावशीलता को कवर करता है, सिर्फ मंजूरी नहीं
समाधान चरण पेप्टाइड संश्लेषण पेप्टाइड आपूर्तिकर्ता परिवर्तन नियंत्रण के अंतर्गत 2026 संस्करण हस्ताक्षरित अनुमोदन से अधिक मांगता है. संशोधित पाठ का एक खंड-स्तरीय पूर्वाभ्यास खंड पढ़ता है 6.3, परिवर्तन की योजना, सूचना की उपलब्धता की अपेक्षा के अनुरूप, परिवर्तन का संचार कैसे किया जाता है, प्रभावशीलता की निगरानी और मूल्यांकन कैसे किया जाता है, और परिणामों की समीक्षा कैसे की जाती है (एमओएल परिवर्तन, 2026-09-29). अकेले अनुमोदन से अब कोई शर्त लागू नहीं होती.
पेप्टाइड क्रो यह मायने रखता है क्योंकि एक राल प्रतिस्थापन, एक काउंटरियन एक्सचेंज या एक विधि-संस्करण परिवर्तन पहचान को बदल सकता है, पवित्रता, अशुद्धता प्रोफ़ाइल या स्थिरता जबकि विश्लेषण का प्रमाण पत्र अपरिवर्तित दिखता है. धारा 8.5.6, परिवर्तनों का नियंत्रण, के बाद से आवश्यक है 2015 आधारभूत बात यह है कि अनपेक्षित परिणामों को रोकने के लिए उत्पादन परिवर्तनों की समीक्षा और नियंत्रण किया जाना चाहिए; अपेक्षित कलाकृतियाँ परिवर्तन अनुरोध प्रपत्र हैं, जोख़िम का आकलन, अनुमोदन रिकॉर्ड और अद्यतन प्रक्रियाएँ (क्यूए, फ़रवरी 2026).
इसलिए पिछले से एक विशिष्ट परिवर्तन के पीछे परिवर्तन रिकॉर्ड के लिए पूछें 24 महीने और अंत से अंत तक चलते रहें: अनुरोध, जोखिम आकलन, अनुमोदन, प्रभावित ग्राहकों से संचार, प्रभावशीलता की जाँच, बंद. अनुमोदन तिथियों वाला एक लॉग और कोई प्रभावशीलता समीक्षा नहीं, या एक ग्राहक अधिसूचना जो शिपमेंट के बाद उतरती है, विफलता मोड है. एक उदाहरण: एचपीएलसी शुद्धता परख को एक नई विधि संस्करण में ले जाया गया, परिवर्तन को मंजूरी दे दी गई, और किसी ने भी एक ही लॉट पर दोनों संस्करणों की तुलना नहीं की.
अभ्यास 2: बैच ट्रैसेबिलिटी आप पीछे और आगे का पुनर्निर्माण कर सकते हैं
पेप्टाइड बैच ट्रैसेबिलिटी एक फाइलिंग अभ्यास नहीं है, और ऑडिट परीक्षण इसे साबित करता है: हाल की डिलीवरी से एक लॉट नंबर चुनें और आपूर्तिकर्ता से इसे फिर से बनाने के लिए कहें, अभिलेखों से, without preparation time. Backward, the genealogy should run to raw-material lots and their suppliers. Forward, it should run to every shipment पेप्टाइड परीक्षण that drew on the lot. The clause 8 operation requirements set the baseline: खंड 8.5.2 requires organisations to identify outputs where needed, identify their status (approved, pending, rejected, पकड़ना), और, where traceability is required, control unique identification and retain the documented information behind it.
That is why a certificate of analysis alone proves less than buyers assume. A CoA reports results for a sample; it does not show which raw-material lots entered the batch, which method version was run, or where the rest of the lot went. Traceability is the artifact that tells you whether the certificate means anything.
टिप के लिए: Choose the lot number before the call, not during it. A supplier who knows the sample in advance can assemble a tidy narrative; a supplier with live records answers in minutes. दुकान
क्या 2026 edition expects
Artifact to request
धारा Semaglutide Synthesis anchor
Outputs identified, status visible, unique identification controlled where traceability is required, documented information retained
A backward-to-materials and forward-to-shipment reconstruction of one sampled lot, produced from records Liquid Phase Peptide Synthesis
धारा 8.5.2, identification and traceability
Failure looks like one of two things. Either the genealogy stops at the bulk intermediate, leaving the raw-material lots unnamed, or the reconstruction takes days and arrives as a written account rather than as records. Both tell you the same thing: the system holds a certificate, not a traceable lot.
अभ्यास 3: मूल कारण और सीएपीए लिंक के साथ विचलन प्रबंधन
Deviation management in peptide manufacturing is where a supplier’s quality system either works or performs. The test is not whether deviations are closed, but whether each one closes with a determined cause and a corrective action whose effectiveness was reviewed.
The clause 10.2 requirements in full set the sequence to check each entry against: react to the nonconformity by controlling and correcting it and dealing with its consequences; evaluate whether action is needed to eliminate the cause so it does not recur or occur elsewhere; determine the causes; determine whether similar nonconformities exist; implement the action; review its effectiveness; update risks and opportunities; and change the quality management system if necessary. The retained record must show the nature of the nonconformity and the actions taken, including results.
Request the last 12 months of the deviation log and read it for three things. Are causes determined, or merely described? Was the rest of the operation checked for the same nonconformity? Was effectiveness reviewed after implementation?
A log where every entry reads “operator error, retrained” fails all three. An isolated deviation is a data point; the same one recurring across products, instruments or operators is a systemic signal, and only the CAPA record separates them.
अभ्यास 4: दस्तावेज़ीकरण और विश्लेषणात्मक रिकॉर्ड जो निरीक्षण में जीवित रहते हैं
The documentation question is not whether records exist. It is whether they are controlled, attributable and retained in a form an inspector can follow. A supplier can hand you a certificate of analysis and a tidy summary table and still fail this test, because a summary table is a transcription of a result, not the record that produced it.
Start with the controlled document set index: the procedures, the method versions and the specifications, each with a version number and an effective date. A method referenced by name with no version is a gap, विवरण नहीं. Then ask for the raw data behind one reported purity or identity result: the chromatogram and the mass spectrum, with the acquisition method version, the analyst and the instrument identified.
That request maps to the data-integrity principles set out in MHRA’s data-integrity guidance: कारण, पढ़ने योग्य, समसामयिक, मूल, शुद्ध. The documented-information requirement in clause 8 of ISO 9001:2026 उसी तरह इंगित करता है, toward records that are created and controlled as the work happens rather than reconstructed afterward.
⚠️ चेतावनी: A summary table is not raw data. If the package contains a CoA and a table but no chromatogram, you cannot verify the result, only accept it.
When a supplier calls the raw data proprietary, that is a commercial position, not a quality one. Ask instead for a redacted chromatogram with the method version and instrument intact. The result and its provenance are what you are evaluating; the formula behind a proprietary gradient is not.
अभ्यास 5: डिज़ाइन द्वारा गुणवत्ता और तुलनीयता वार्तालाप
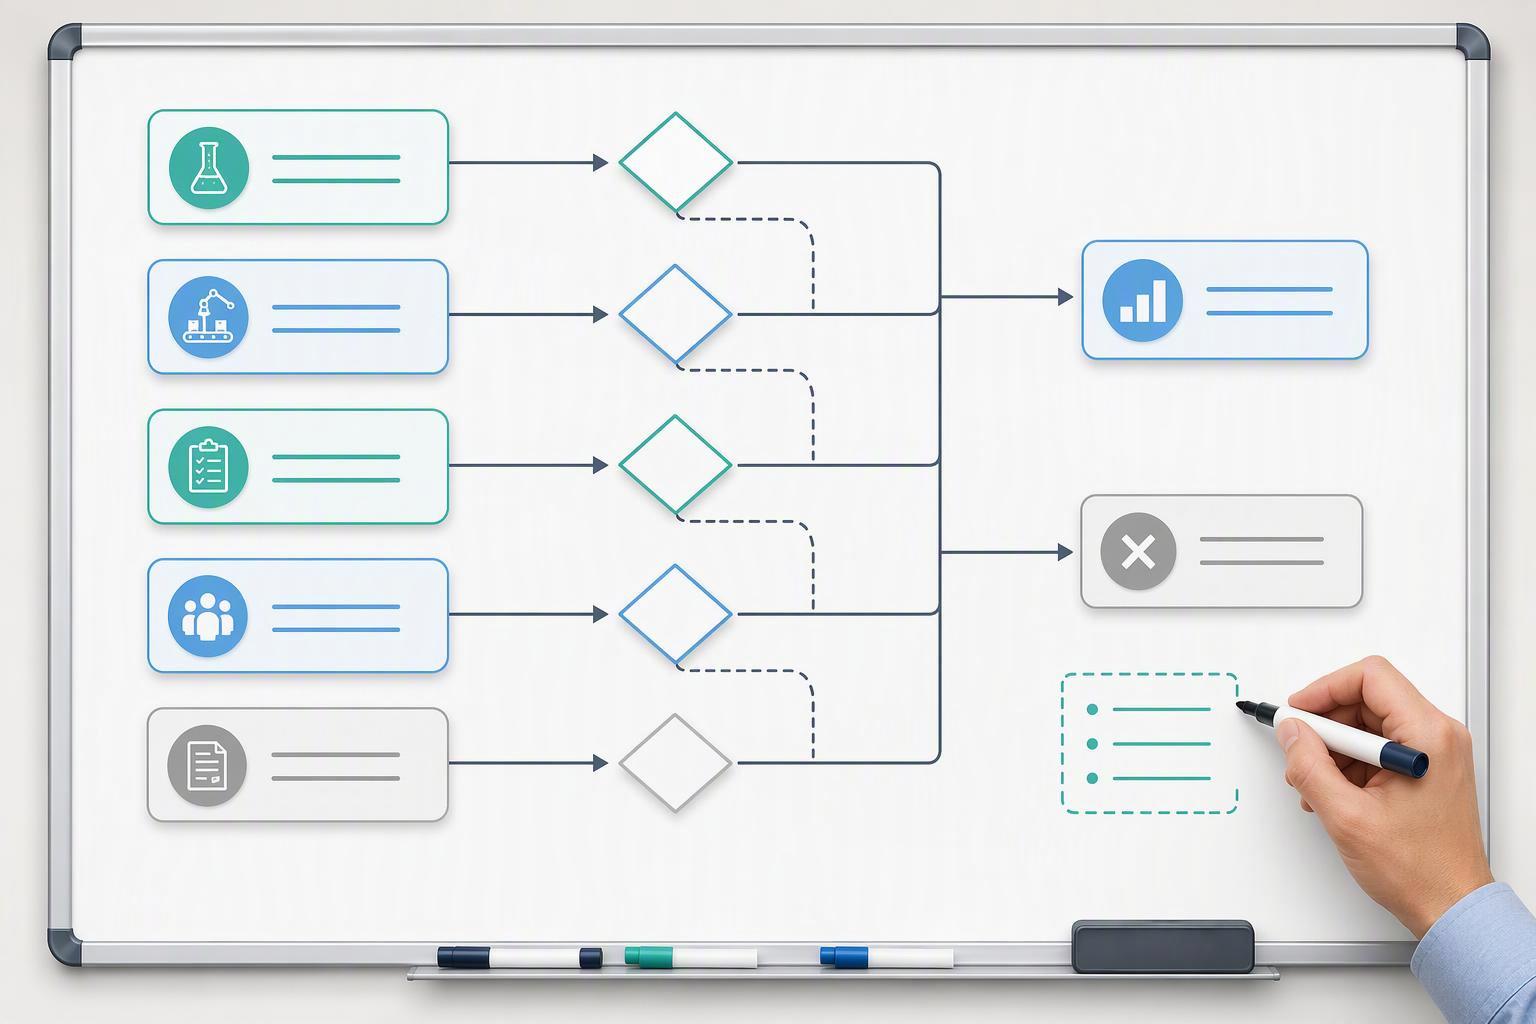
Quality by design for a peptide supplier means one practical thing at the buyer’s end: a comparability protocol agreed before a change, not a discussion after it. At peptide scale, purity and yield move together, so a change that looks operationally minor can shift the impurity profile.
That trade-off is why scale-up changes carry risk. CDMO guidance on milligram-to-kilogram scale-up notes that above 95% purity at kilogram scale the route often necessitates multi-step preparative HPLC and carefully controlled lyophilization, and that aggregation, निम्नीकरण, or prolonged hold times can cause notable yield loss. The same source puts the practical solid-phase limit near 30 को 40 अवशेष. It is a CDMO’s own explainer, so read it for direction rather than as an independent benchmark.
Synthesis economics reinforce the point. The widely cited synthesis-economics analysis puts a 50-plus-step process at an average 99% yield per step, and reports that improving crude purity by 10% can save more than 50% post-purification. The figures come from named industry experts quoted in that piece, not from a controlled study.
Put the protocol in the quality agreement: which change types trigger a comparability study, what it measures (पहचान, पवित्रता, अशुद्धता प्रोफाइल, and where relevant potency and stability), and what acceptance criteria apply before changed material ships. A supplier that reports a change after the fact and offers a CoA as the comparability evidence has not met that bar.
A documentation package that includes full analytical verification, such as the one MOL Changes can be used to provide, supports this review.
इसे लागू करते समय खरीदार सामान्य गलतियाँ करते हैं 2026 अपेक्षाएं
The most common buyer-side failure is treating the certificate as the evidence rather than the records behind it. A certificate says an audit happened; it does not show that the supplier can reconstruct a batch, explain a deviation, or control a change. These five mistakes show up repeatedly in supplier evaluations, and each has a straightforward fix.
Accepting a CoA as proof of traceability. A certificate of analysis confirms what was tested on one sample, not where the material came from or where it went. Buyers accept it because it arrives first and looks authoritative. Ask instead for the batch genealogy that links incoming lots to the finished peptide.
Reading a change log for approvals only. A log showing sign-offs tells you a change was authorized, not that anyone checked whether it worked. The fix is to request the effectiveness check that follows the approval.
Accepting a deviation log with dispositions and no causes. A disposition closes a batch; a root cause and a linked CAPA close the problem. If the log has no cause column, the supplier is tracking outcomes rather than preventing recurrence.
Requesting a documentation package without naming the artifacts. Vague requests get vague packages. Name what you want: the change record, the deviation log with CAPA references, the controlled document set.
Leaving the transition obligation out of the quality agreement. If the agreement is silent on who owns the 2026 transition, the obligation defaults to nobody. Put the deadline and the responsible party in writing.
अक्सर पूछे जाने वाले प्रश्नों
What happens to a supplier’s 2015 प्रमाण पत्र के बाद 30 सितम्बर 2029?
It stops being a valid basis for your qualification file. Under the ISO 9001:2026 transition timeline, certificates issued against the 2015 edition are withdrawn on 30 सितम्बर 2029 (the ISO/TC 176 transition notice). If a supplier has not transitioned by then, treat their certificate as expired and re-open the qualification rather than letting it lapse quietly in your approved-supplier list.
क्या किसी आपूर्तिकर्ता को प्रकाशन के तुरंत बाद पुनः प्रमाणित करना होगा?
नहीं. आईएसओ 9001:2026 पर प्रकाशित किया गया था 16 सितम्बर 2026 (ISO’s launch announcement), and the transition window runs to 30 सितम्बर 2029. A supplier holding a current 2015 certificate stays certified during that period. What changes for you is the deadline, not the certificate: ask for their transition plan and a target audit date, then track it as a qualification milestone.
करता है 2026 संस्करण आपूर्तिकर्ता-योग्यता आवश्यकताओं को स्वयं बदलता है?
Not the requirement to evaluate and control externally provided processes, products and services, which remains part of the standard’s purchasing and supplier control expectations. क्या 2026 edition shifts is how you evidence that evaluation: नियंत्रण बदलें, पता लगाने की क्षमता, deviation handling and documented records are assessed against the practices above. Your qualification criteria stay yours; the audit trail behind them gets stricter.
यदि कोई आपूर्तिकर्ता ऑन-साइट ऑडिट से इनकार कर दे तो क्या होगा??
Accept it as a data point and adjust the evidence you require. Ask instead for a remote documentation review, a completed self-assessment against the five practices, batch records for a named lot, and a deviation log with CAPA linkage. If they decline all of these, you have no verifiable basis for approval, and the practical answer is to keep them unapproved or qualify an alternative source.
निष्कर्ष
You now have five artifacts to request from any peptide supplier you are qualifying: a change-control record that shows communication and effectiveness, a batch genealogy you can walk backward and forward, a deviation log linked to a root cause and a CAPA, a controlled documentation and analytical record set, and a comparability protocol for process changes. Each one is a document a supplier either has or does not, which makes the request itself a fast filter. The dates make it timely rather than theoretical: the ISO/TC 176 transition notice sets 30 सितम्बर 2029 as the end of the transition, so a supplier who cannot produce these records now has roughly three years to build them, and you carry the audit risk in the meantime.
If you would rather not run that review alone, talk to a technical expert about your documentation package, or request a walk-through of the change-control and traceability records.
खुलासा: this article is published by MOL Changes, a peptide manufacturer, so the supplier practices described here reflect the standards we are held to as well as the ones we apply.