Como escolher um parceiro Peptide CDMO: Comece com seu próprio mapa de requisitos
A maioria das equipes começa construindo uma lista de fornecedores. A atitude mais forte é anotar primeiro suas próprias restrições, porque um parceiro que parece excelente em uma página de recursos ainda pode ser a opção errada para sua sequência. Cinco entradas fazem o trabalho: classe e comprimento da sequência, tipos de modificação, escala e rota alvo, caminho de arquivamento e cronograma, e os requisitos analíticos e de documentação de peptídeos que sua submissão terá.
A questão do caminho pertence ao topo da lista. Os peptídeos ficam entre pequenas moléculas e produtos biológicos, normalmente regido pelas diretrizes de moléculas pequenas do ICH, embora algumas aulas possam exigir supervisão biológica, é por isso que o alinhamento regulatório precoce é importante (Laboratórios Neuland, 2025). Responda antes de entrar em contato com alguém, já que o caminho determina quais artefatos um parceiro deve ser capaz de produzir.
|
Exigência |
Por que isso restringe a escolha do parceiro |
Artefato que prova isso |
|---|---|---|
|
Classe e duração da sequência |
Determina se a síntese em fase sólida, fermentação, ou uma rota híbrida é viável |
Justificativa de rota com sequências precedentes |
|
Tipos de modificação |
Resíduos não padronizados, conjugados, e ciclização precisam de capacidade química dedicada |
Menu de modificação com cobertura de grupo funcional |
|
Escala e rota alvo |
Lacunas entre desenvolvimento e escala comercial forçam uma transferência de tecnologia |
Histórico documentado de aumento de escala em sua faixa-alvo |
|
Síntese de Peptídeos Caminho de arquivamento e cronograma |
A supervisão de moléculas pequenas do ICH versus a supervisão biológica altera o pacote de evidências |
Avaliação de lacunas regulatórias para o seu caminho |
|
Requisitos analíticos e de documentação |
Os testes de liberação e o escopo de estabilidade devem corresponder ao arquivamento |
Exemplo de pacote de lançamento e resumos de métodos |
Principal vantagem: Preencha este mapa antes de sua primeira ligação com o fornecedor. Ele converte uma pesquisa vaga em uma lista pontuada, e informa quais artefatos solicitar, em vez de quais afirmações acreditar.
Ajuste Técnico: Combinando a plataforma de síntese com sua classe de sequência
Com esse mapa em mãos, o primeiro filtro é a compatibilidade de rota, não volume do reator. Anunciado capacidade específica da rota os números raramente mencionam a rota de fabricação por trás deles, e a capacidade não é transferida entre rotas: fase sólida (SPSS), fase líquida (LPPS), convergente híbrido, e plataformas recombinantes mais sintéticas não são intercambiáveis. Um fornecedor com toneladas sobressalentes em uma linha de LPPS não pode absorver seu trabalho de SPPS.
O comprimento da cadeia decide quais rotas são viáveis. Neuland Labs observa que os rendimentos do SPPS diminuem à medida que as cadeias crescem, com sequências além 30 para 40 aminoácidos aproximando-se do limite prático apenas para SPPS; a condensação híbrida de fragmentos de fase sólida/líquida reduz os custos e mantém a pureza. A matemática composta da eficiência do acoplamento explica por que: no 99% acoplamento por etapa acima 30 passos, apenas 74% de cadeias completas saem corretas, e a dificuldade de expansão se intensifica além 30 resíduos, que é exatamente onde a semaglutida (31) e tirzepatida (39) sentar (Peptide Journal, 2026-02-09).
Então, quando você pergunta como escolher um parceiro peptídico CDMO, a resposta começa com três perguntas: qual rota você propõe para esta sequência, qual estratégia de fragmento essa rota usa, e posso ver os dados de pureza bruta por trás da escolha?
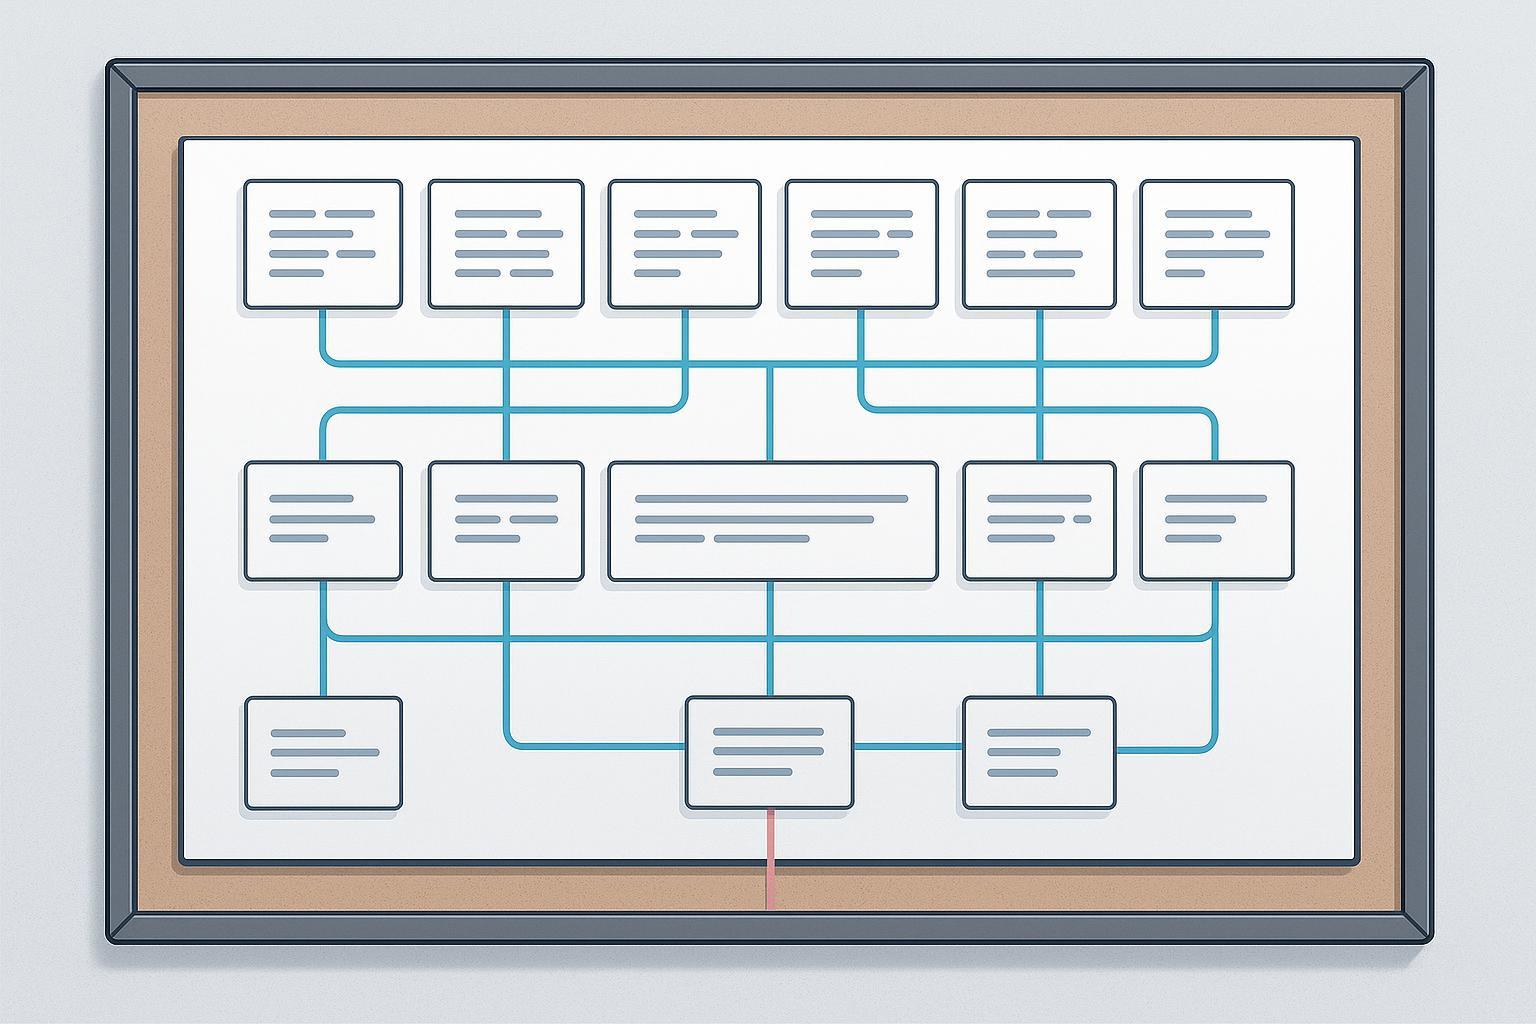
Classe de sequência para mapa de decisão de rota candidata, mostrando peptídeos lineares curtos, cadeias longas ou hidrofóbicas, e sequências modificadas ramificando-se para SPPS, convergente híbrido, LPPS, ou caminhos recombinantes mais sintéticos.
Disciplina de expansão: Verifique o caminho, Não é o tamanho do reator
Compatibilidade de rota resolvida, o tamanho do reator ainda não é um plano de expansão. The binding constraint in most peptide programs is purification, not chain assembly: preparative HPLC accounts for 20 para 35 percent of total peptide manufacturing cost, and a two-week synthesis campaign can require four to six weeks of purification work (PeptídeoStaff, retrieved 2026-06-01). CDMO Hub’s analysis of peptide manufacturing puts the same point bluntly: synthesis capacity is not purification capacity, and for complex or high-volume sequences the preparative HPLC train and lyophilization suite are what actually set the schedule (Issue 4, retrieved 2026-08-26).
That gap widens as peptide scale-up from mg to kg proceeds, because a single GMP-scale preparative HPLC run consumes 20 para 50 liters of acetonitrile and 4 para 12 hours of instrument time (PeptídeoStaff, retrieved 2026-06-01). Solvent logistics, waste handling, and lyophilization throughput therefore deserve the same scrutiny as reactor volume.
A disciplined partner should already hold the high-risk parameters in a defined design space. PharmaFocus America’s QbD review classifies cleavage and deprotection pH and reaction time, HPLC solvent gradient and flow rate, and SPPS coupling efficiency and resin swelling as high-risk parameters requiring explicit control, with design and control spaces fixed by the end of clinical development (Quality by Design in Action, 2025-06-16).
Para dica: Ask three questions before accepting any capacity claim. What is the purification train, including column dimensions and gradient capability? What is the lyophilization suite capacity, in shelf area and cycles per week? And how are manufacturing slots defined in the agreement, by calendar window or by confirmed campaign start?
Profundidade Analítica: O pacote de lançamento sob demanda
Ask for the method list before you ask for the price. A vendor that can name its chiral method, its counterion assay and its endotoxin calculation is describing a release package; one that answers with a purity percentage is describing a brochure.
Every analytical procedure behind that package should be validated under Eu Q2(R2), adopted in March 2024, which sets the general framework for validation, including spectroscopic methods, and covers post-approval change management.
Endotoxin is where loose language shows up fastest. USP <85> sets no single parenteral limit: it is the K/M endotoxin limit, with K = 5 USP-EU/kg for routes other than intrathecal and 0.2 USP-EU/kg for intrathecal, and M as the maximum recommended human dose per kg per hour. A vendor quoting an absolute EU/mg figure without the dose basis has not done the calculation.
Sterility follows the same pattern. The two USP sterility methods are membrane filtration and direct inoculation, both requiring 14-day incubation, with Fluid Thioglycollate Medium at 30-35 °C and Soybean-Casein Digest Medium at 20-25 °C. Confirm which method is used and on what sample size.
Impurity control is the criterion that separates in-process discipline from end-only testing. The main peptide impurity classes are deletion sequences, incomplete deprotection products, over-coupled and side-reaction products, produtos de oxidação, and residual reagents, solvents and counterions. In-process analytics, UV monitoring for Fmoc removal, Kaiser or ninhydrin testing for coupling completeness, and HPLC/MS on crude and purified material, catch these while the batch is still recoverable.
⚠️ Aviso: A high HPLC area percent is not identity confirmation, and a correct intact mass is not a purity measurement. ICH Q6A notes that a single chromatographic retention time is not sufficiently specific, and counterion content such as TFA, acetate or chloride does not appear in an HPLC-UV peptide purity figure at all.
The thresholds worth writing into your requirements map come from the 0.10% e 0.5% impurity thresholds in FDA’s 2021 orientação: impurezas relacionadas a peptídeos em 0.10% or greater should be identified and characterised, new impurities between 0.10% e 0.5% need characterisation plus justification including a comparative immunogenicity risk assessment, and new impurities above 0.5% of drug substance are not acceptable for that ANDA pathway.
Use the table below as the checklist you send with your RFP. Each row names a test, the standard it rests on, and the artifact the vendor should hand over so you can verify the claim rather than accept it.
|
Teste |
Standard or basis |
Artifact the vendor supplies |
|---|---|---|
|
Chiral purity |
Validated chiral method under ICH Q2(R2) |
Representative chromatogram plus validation summary |
|
Identidade e conteúdo do contra-íon |
Serviços ICH Q6A specificity Peptídeos Sintéticos expectativas |
Assay method and result for TFA, acetate or chloride |
|
Solventes residuais |
ICH Q3C limits |
Headspace GC method and batch results |
|
Endotoxina |
USP <85>, K/M calculation |
LAL result with the dose basis used for M |
|
Esterilidade |
USP <71>, membrane filtration or direct inoculation |
Método, incubation conditions, 14-day result |
|
Peptide-related impurities |
FDA 2021 limites de impureza |
Impurity table with identification and characterisation data |
|
Identidade |
Eu Q6A |
Mass spectrum plus a second orthogonal identity method |
Treat these as peptide analytical and documentation requirements, not as a wish list. If a vendor cannot produce the artifact for a row, that row is unverified, and unverified analytical depth is the most expensive gap to close later, because it surfaces during filing rather than during the RFP.
Documentação e prontidão regulatória: Arquivando artefatos como pontos de verificação
Documentation is where a peptide CDMO partner either shortens your filing timeline or quietly extends it. Evaluate it as filing artifacts, not as a quality certificate.
Request the analytical method documentation first, and check it against the ICH Q2(R2) validation framework: especificidade, precisão, precisão, linearidade, range, and robustness each need a stated result, not a claim of compliance. Then ask for the impurity rationale. The impurity-rationale expectation is a documented justification for every specified and unspecified impurity, tied to the route of synthesis and to forced-degradation data. A profile with no rationale behind its limits is a filing risk you inherit. Produção de Peptídeos
Read batch records through the ALCOA data-integrity lens set out in MHRA’s GXP data integrity guidance, then test it with one question: how is a deviation recorded, investigated, and closed? The answer reveals whether change history is a controlled record or a retrospective summary.
Put four items in the technical agreement: registros em lote, justificativa de impureza, change history with effective dates, and the DMF or ASMF position. Add the supplier-tier audit gap: sponsors audit the CDMO but not its input suppliers, and final-assembly equipment lead times are reported at 18-24 meses (CDMO Hub, Por Dentro da Fabricação de Peptídeos, Issue 4, 2026).
Modificações especializadas sem atrasos na transferência
Every outsourced modification step is a handoff, and handoffs are where timelines are lost. The modification types that most often leave the primary synthesis site are lipidation, PEGuilação, conjugação, non-natural and D-amino acid incorporation, ciclização, and counterion exchange.
The cost of an unplanned handoff is measurable in planning terms. Qualifying a new peptide API supplier is typically quoted at six to twelve months, with the full second-source path commonly cited at eighteen to twenty-four months (CDMO Hub, “Six Peptide CDMO Selection Pitfalls,” retrieved 2026-08-26). These are directional planning ranges from vendor-adjacent trade publishers, not audited figures, so treat them as a scheduling assumption rather than a benchmark.
A subcontracted step can also sit on a different route and a different capacity pool than the main chain, which means the modification queue moves on someone else’s schedule. That is why the modification-capability question belongs in the technical evaluation: an integrated partner should hold in-house cyclic, PEGylated, grampeado, conjugated and unusual-amino-acid capability (Laboratórios Neuland, retrieved 2026-07-16).
One practical way to confirm specialized peptide modification support stays inside the same system is a verification workflow rather than a capability claim. Ask the sponsor-side technical lead to trace one modification step end to end: which change-control record governs it, which analytical release specification applies, who signs the deviation, and whether the certificate of analysis is issued under the same quality system as the main chain. If the answers route through a third party’s quality department, the handoff is real regardless of how the proposal describes it. MOL Changes is one example of a platform that keeps synthesis and modification under a single accountable workflow.
The contract checkpoint follows from that trace. Name the modification steps explicitly in the quality agreement, require the same change-control and release system for each, and set a notification obligation before any step is moved to another site.
Bandeiras vermelhas e quebra de acordos: O que deve encerrar a conversa
Red flags are not the same as missing must-haves. A missing must-have is a gap you can ask a vendor to close; a red flag is a pattern that tells you the vendor is answering from a brochure rather than from your sequence. Score the two lists separately, because a candidate can pass every must-have on paper and still fail on how the answers were produced.
The two structural red flags both concern capacity claims. A vendor that asserts capacity without naming the route it applies to is describing a general capability, not your project, because route-specific capacity does not transfer across synthesis routes. The same applies to the synthesis-versus-purification gap: synthesis capacity is not purification capacity, so a reactor count tells you nothing about whether the purification train can handle your load.
The remaining deal-breakers are documentation failures. A release package that stops at appearance and assay has no impurity rationale, and at filing the threshold is unforgiving: new impurities above 0.5% of drug substance are not acceptable for that ANDA pathway. Refusal to show a change-history log, a subcontracted modification step with no named accountable owner, and endotoxin or sterility testing described without the applicable limit or method all belong on the same list. So does any purity claim that leans on HPLC-UV alone, which is why identity and purity are different tests.
Pontuando candidatos de acordo com seus próprios requisitos
Turn the requirements map into a weighted matrix before you read another proposal. List each criterion from your opening map, assign it a weight that sums to 100 across the set, then score every candidate from 1 para 5 on the artifact they actually supplied. An absent artifact scores zero, not a neutral three: an unreturned chromatogram is missing evidence, not a rounding error.
Two rows deserve their own weight columns. The first is total-engagement cost, because a quoted price is not the cost. Neuland estimates that an RFP price may represent only 60 para 70 percent of true engagement cost once tech transfer, desenvolvimento analítico, change orders and stability storage are added, which is a single-company estimate rather than an industry benchmark, so treat it as a prompt to build your own line items. The second is slot certainty. PeptideStaff’s mid-2026 US utilization estimates put US peptide CDMO capacity utilization at 78 para 85 por cento, with Phase 1 supply near 82 percent and Phase 2 para 3 near 78 percent at Tier 1 CDMOs, and commercial capacity described as tight; the analyst behind the figure is unnamed, so confirm current availability directly rather than relying on the range alone.
|
Critério |
Peso |
Candidate A |
Candidate B |
Candidate C |
|---|---|---|---|---|
|
Sequence-class fit |
||||
|
Scale-up path evidence |
||||
|
Release package completeness |
||||
|
Regulatory artifact readiness |
||||
|
Modification coverage |
||||
|
Total-engagement cost |
||||
|
Slot certainty |
||||
|
Weighted total |
100 |
Score each cell only against documentation you have in hand, and leave a cell blank rather than guessing. The matrix is a comparison tool, não é um veredicto: it shows you where two candidates genuinely differ and where the difference is only in how well they answered the RFP.
Próximas etapas: Transformando a estrutura em uma conversa técnica
The requirements map, the artifact checklist, and the scored shortlist are only useful once they become one specific request. Send the shortlisted CDMO a single document that pairs your sequence class, scale target, and modification list with the exact artifacts you need to see: method package, analytical documentation, and a defined slot commitment. That request is answerable. A general capabilities deck is not.
Ask for the method package and the analytical documentation before you ask for pricing. Pricing without a method behind it cannot be compared across candidates, and the comparison is the whole point of the framework above. Sobre
For a worked sense of how to choose a peptide CDMO partner, the closing move is the same at every scale: convert the framework into one technical conversation, then let the artifacts decide.
Observação: MOL Changes is a peptide CDMO and may be one of the candidates you evaluate. This article is educational and does not recommend any single supplier.
Readers should consult qualified regulatory and quality professionals before making filing or sourcing decisions.
Perguntas frequentes
Quanto tempo leva para qualificar um parceiro de peptídeo CDMO?
Trade-publisher planning ranges put a Phase 1 new program start at 9 para 15 meses, up from 6 para 9 months in 2023, with commercial manufacturing or tech-transfer starts at 12 para 18 months and a full second-source qualification path commonly cited at 18 para 24 meses. These are planning ranges reported by trade publications, not audited figures, so treat them as directional. The practical implication is the 12-to-18-month engagement rule: begin qualification well before you need GMP synthesis, not after your clinical timeline is already fixed.
Can a CDMO’s announced capacity be used for my sequence?
Not by default. Announced capacity figures rarely specify the manufacturing route they were built for, and route-specific capacity does not transfer between routes, so a headline number tells you little about your sequence. Synthesis capacity is also not purification capacity, and the synthesis-versus-purification gap is where most assumptions break. Two questions resolve it: which route is the announced capacity built for, and what purification and lyophilization capacity accompanies it.
O que devo fazer se uma etapa de modificação tiver que ser subcontratada?
Treat the subcontract as a controlled handoff rather than an exception. Four checkpoints make it manageable: a named accountable owner on the prime CDMO’s side, the same change-control system covering both parties, a defined analytical release point at the handoff, and a documented transfer plan with acceptance criteria. Handoff delays are a recognized risk in multi-site programs, though published figures on their frequency are single-source and should not be treated as a failure rate.
A resposta RFP mais barata é o menor custo total?
Não. Neuland estimates that an RFP price may represent only 60 para 70% of true engagement cost once tech transfer, desenvolvimento analítico, change orders, and stability storage are added, and that figure comes from a single company rather than an independent benchmark. Score responses on the total-engagement-cost row of your matrix, not the quoted line item, and ask each candidate to price the scope you actually expect to consume.
Conclusão
The framework reduces to four moves you can reuse on any candidate: map your own requirements before you take a call, ask for artifacts rather than assertions, score red flags separately from must-haves, and put total-engagement cost into the matrix instead of comparing headline quotes.
The market context makes that discipline worth the effort. With mid-2026 US utilization estimates at 78-85%, announced capacity is not the same as capacity you can book, and lead times still stretch well past the point where a program can absorb a restart. A partner who can show you the synthesis route, the analytical package, and the change history is worth more than one who can only show you a reactor count.
Como o 2024-2026 capex cycle’s capacity comes online through 2027-2030, the differentiator shifts from who has capacity to who can document it. The conversation you start now, framed around your requirements map, is the one that will still be useful when the supply picture loosens.