O que a autorização ENDO-205 diz ou não sobre as expectativas pré-IND do peptídeo CMC
A ação da FDA do ENDO-205 foi uma autorização do IND, não é uma aprovação, e não diz nada sobre o pacote CMC por trás disso. Marcha da EndoCyclic 2026 Anúncio de liberação do IND descreve um peptídeo direcionado não hormonal de primeira classe para endometriose, namorou Irvine, Califórnia., Marchar 23, 2026, com uma fase 1 estudo planejado em mulheres saudáveis na pré-menopausa em idade reprodutiva. O mesmo comunicado afirma que o trabalho pré-clínico mostrou eliminação de lesões de endometriose e inflamação associada sem sinais de segurança em estudos de toxicologia GLP. Não contém CMC, fabricação, CDMO, ou detalhe de síntese de qualquer tipo.
Essa lacuna é o ponto. Um IND liberado informa que um pacote pode ser revisado o suficiente para prosseguir; não diz o que havia nele, e não é um modelo que você possa copiar. Uma análise independente do mecanismo do ENDO-205 descreve uma permeação celular, Peptídeo sensível ao pH combinando inibição da via Wnt / β-catenina com entrega sensível ao pH, inerte no pH do tecido saudável e ativo no microambiente da lesão acidificada. Esse detalhe vem apenas da fonte secundária, não do patrocinador, e a seletividade foi demonstrada apenas em modelos pré-clínicos.
⚠️ Aviso: O espelho bioespacial, a listagem da PR Newswire, um item farmacêutico, e um compartilhamento do LinkedIn parafraseiam um lançamento do EndoCyclic. Eles contam como uma fonte, não quatro. Nenhuma fonte cobrindo o desenvolvimento do ENDO-205 após março 2026 foi encontrado, então este artigo trata o evento como uma data histórica fixa.
O restante deste guia trabalha através dos cinco pilares que determinam se os requisitos CMC de um depósito IND de peptídeo são atendidos: confirmação de sequência, perfil de impurezas, qualificação de método analítico, reprodutibilidade do processo, e documentação.
Por que o CMC é o motivo mais comum para a paralisação dos primeiros INDs
Química, a fabricação e os controles são a base mais comum para uma suspensão clínica em um IND precoce. Uma revisão piloto retrospectiva do ano fiscal 2013 A coorte do CDER descobriu que 1,410 INDs iniciais recebidos naquele ano, 125, ou 8.9%, foram colocados em espera no primeiro 30 dias, e as razões da CMC foram citadas com mais frequência do que as razões clínicas ou toxicológicas (o estudo piloto da FDA sobre a clínica IND mantém, publicado 2016). Leia isso como uma linha de base histórica em vez de uma taxa atual: é uma revisão de um ano de uma única coorte, e os padrões de espera mudaram desde.
A mesma descoberta é o que torna o trabalho pré-IND do peptídeo CMC tratável. A FDA afirma que a quantidade de informações necessárias “variará de acordo com a fase da investigação, a formulação proposta, e duração da investigação” (Requisitos de informação IND CMC da FDA, conteúdo atual a partir de 22 Junho 2026). Fase 1 Os requisitos CMC para arquivamento do peptídeo IND são, portanto, um conjunto de escopo, não é o pacote comercial completo.
Os cinco pilares da preparação pré-IND do peptídeo CMC
O pacote pré-IND do peptídeo CMC é reduzido para cinco pilares, e a questão útil não é se cada um deles importa, mas quando deve ser concluído. Os requisitos de informação IND CMC da FDA listam cinco elementos de substâncias medicamentosas que um patrocinador deve eventualmente fornecer: estrutura e evidência de identidade, the manufacturer’s name and address, the general method of preparation with reagents, solvents and catalysts, acceptable limits plus the analytical methods that enforce identity, força, quality and purity, and stability support for storage and for the toxicological and clinical studies (FDA, Aplicações IND: Informações CMC, conteúdo atual a partir de 2026-06-22).
Those five elements map onto the pillars below. What changes between Phase 1 and later phases is depth, not the list itself, so the practical split is what you lock before the meeting versus what you defer with a written rationale.
|
Comprimidos |
Lock before pre-IND |
Can be deferred with documented rationale |
|---|---|---|
|
Sequence confirmation |
Orthogonal confirmation of the intended sequence on the clinical lot |
Extended sequence variant mapping beyond the clinical lot, with a rationale citing lot coverage |
|
Impurity profiling |
Identification and quantitation of peptide-related impurities against the applicable thresholds |
Full toxicological qualification of every specified impurity, with a rationale citing the qualification threshold and exposure |
|
Métodos analíticos |
Qualified methods with defined acceptance criteria for identity, força, quality and purity |
Formal validation protocols, with a rationale citing the phase-appropriateness of qualification at this stage |
|
Process reproducibility |
Lot-to-lot comparability data across the lots used in toxicology and clinical supply |
Serviços Formal process validation and continued process verification, with a rationale citing the number of lots manufactured to date |
|
Documentação |
The CMC narrative and the written pre-IND question set |
The full Module 3 dossier, with a rationale citing the pre-IND stage |
The deferral column is the part sponsors under-document. Each deferred item needs the rationale recorded in the briefing package itself, not held in internal notes, because the reviewer’s question at the meeting is usually why a given piece of data is absent rather than whether it exists.
Comprimidos 1: Confirmação da sequência antes da reunião pré-IND
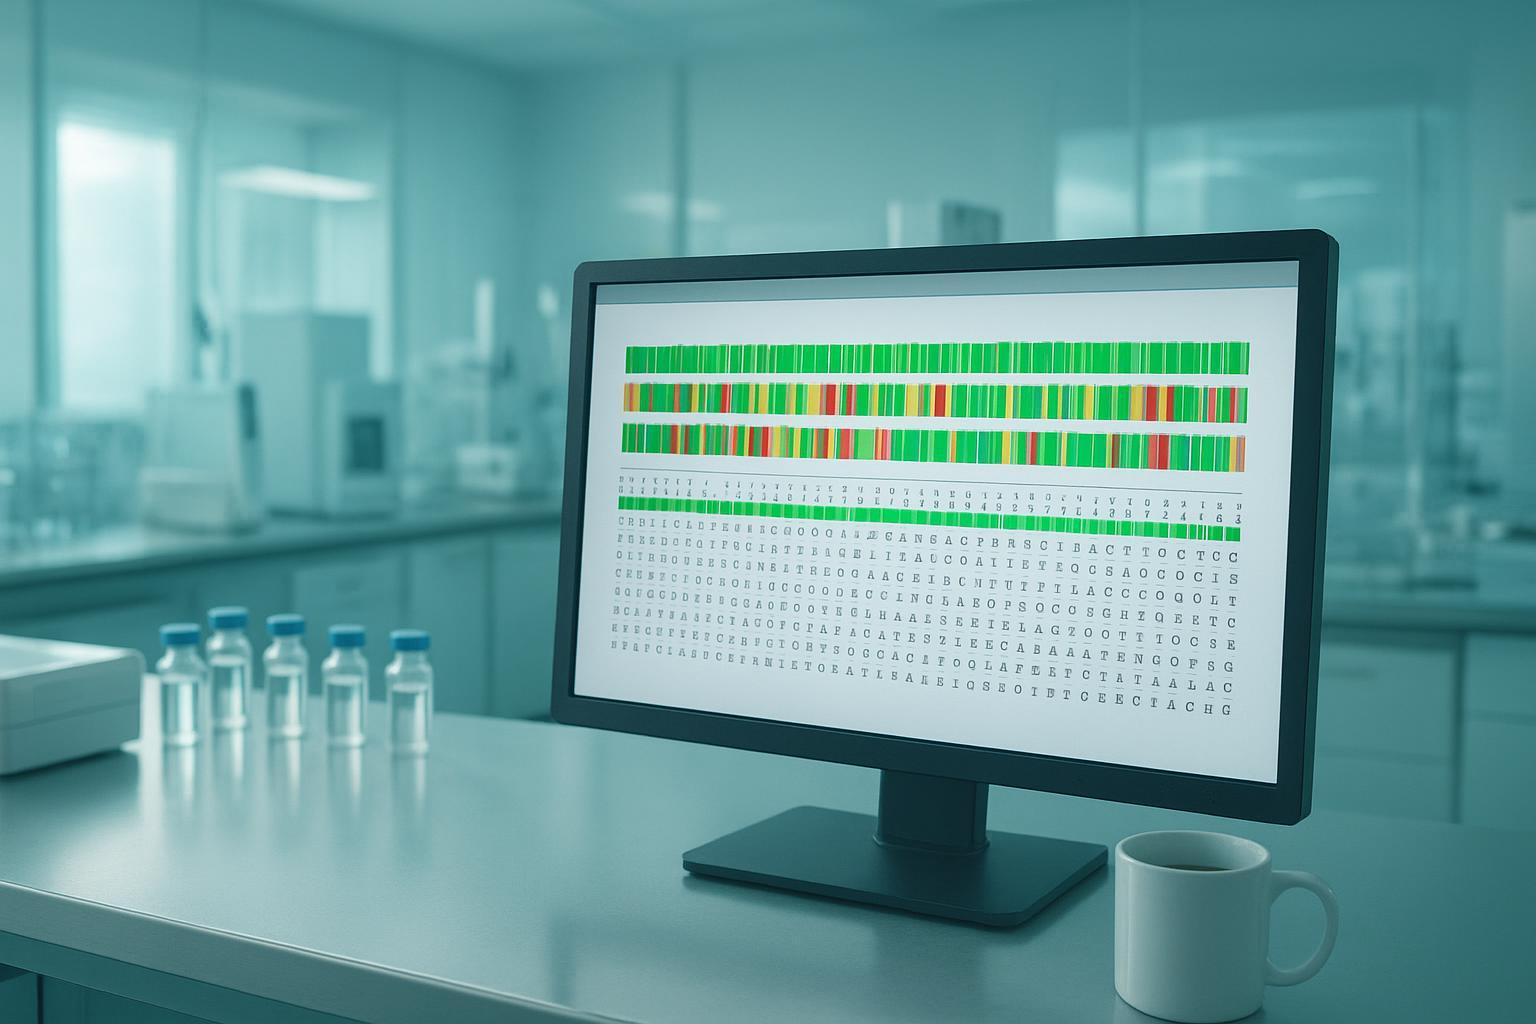
By the end of this pillar you should hold a confirmed primary sequence tied to a specific lot, not a certificate of analysis alone. FDA’s IND CMC information requirements call for a description of the drug substance’s physical, químico, or biological characteristics plus evidence supporting its structure and identity (FDA, recuperado 2026-06-22). Note what that page does not do: it never names “amino acid sequence” or “peptide map,” so build your peptide sequence confirmation on the peptide-specific guidance instead. FDA’s acceptance criteria for synthetic peptides treat primary sequence as one of several sameness dimensions alongside physicochemical properties, secondary structure, oligomer and aggregation states, and biological activity (RAPS).
The workflow that satisfies this: intact mass by high-resolution mass spectrometry, then enzymatic or chemical fragmentation with LC-MS/MS to establish sequence coverage across the full chain. Where coverage gaps remain, confirm those regions orthogonally. Lock the reference standard against the confirmed mass profile so every later lot is compared to a characterized anchor rather than to a vendor’s summary.
⚠️ Aviso: A sequence confirmed only by a vendor CoA, with no traceable raw method output behind Comprar it, is the failure mode. If you cannot produce the coverage map, you have not confirmed the sequence.
Comprimidos 2: Perfil de impurezas e os limites que realmente se aplicam aos peptídeos
Peptides are explicitly excluded from ICH Q3A. The Federal Register notice reproducing the guidance lists the drug substance types it does not cover, and peptide sits in that list alongside biological products, oligonucleotides, and radiopharmaceuticals (ICH Q3A’s scope exclusion for peptides, 2003-02-11). The practical consequence is that the Q3A reporting, identificação, and qualification table is a reference framework for a peptide program, not a binding requirement, and peptide impurity control is risk-based.
FDA’s own rationale for peptide-specific criteria explains why. As quoted by RAPS from the final guidance, differences in impurities, particularly peptide-related impurities, may affect the safety or effectiveness of a peptide drug product compared with the reference listed drug (FDA’s acceptance criteria for synthetic peptides). That reasoning drives the numbers a sponsor should plan around: a specified peptide-related impurity at no more than 0.5% of the drug substance, and each new specified peptide-related impurity specified with justification for why it would affect safety or effectiveness versus the reference listed drug, with each impurity present at equal or lower levels in the generic (FDA’s acceptance criteria for synthetic peptides).
The ICH Q3A thresholds themselves are dose-band dependent, and the framework is best treated qualitatively here: the reporting and identification levels that apply to a small molecule shift with maximum daily dose, so the same percentage does not carry the same meaning across programs (the ICH Q3A reporting and identification thresholds, Q3A(R2), 2006). The failure mode is specific and common: quoting a 0.10% identification threshold as if it bound a peptide program, then defending that number at the pre-IND meeting when the reviewer asks why the control strategy was built on a guideline that excludes the molecule.
Comprimidos 3: Métodos Analíticos — Qualificação Agora, Validação mais tarde
At Phase 1, you need qualified methods with documented suitability, not fully validated ones. The distinction is explicit in FDA’s Phase 1 CMC flexibilities: the agency expects acceptable limits and analytical methods to ensure identity, força, quality and purity, with a brief description of the test methods used, as set out in FDA’s IND CMC information requirements. Full validation under ICH Q2(R2) can follow.
A peptide program typically needs RP-HPLC or UHPLC purity, LC-MS/MS and HRMS for identity and impurity structure, CE, análise de aminoácidos, counterion and TFA content, solventes residuais, teor de água, and endotoxin by LAL. Anchor validation expectations to ICH Q2(R2); anchor endotoxin methods to USP <71> e USP <85>.
⚠️ Aviso: A purity method with no forced-degradation or specificity data behind it is the failure mode. Qualification without documented suitability is not qualification.
Comprimidos 4: Reprodutibilidade do Processo e Narrativa de Comparabilidade Lote a Lote
Reproducibility is demonstrated by overlaying lot data across at least three lots and showing that the impurity and assay profiles agree, not by asserting that the process is controlled. A sponsor who states “the process is reproducible” without the overlay has made a claim the reviewer cannot check.
The regulatory hook is the general method of preparation. FDA’s IND CMC information requirements ask for a description of the method of preparation including reagents, solvents and catalysts, plus stability information for the intended container closure and for the toxicological and clinical studies. Read together, those two items are a reproducibility argument: the method description explains what should stay constant, and the lot data shows that it did.
Síntese de Peptídeos Fold sequence confirmation into the same narrative. The sequence data establishes what the molecule is; the lot overlay establishes that you can make it again. Presenting them as one comparability argument, rather than two separate attachments, is what closes the gap reviewers most often flag.
The failure mode is specific. Three lots can each pass individually while their impurity overlays diverge at a specified impurity, which tells the reviewer the process is not yet under control even though no single lot breached a limit. O 0.5% specified-impurity limit is a ceiling, not a target, and FDA’s acceptance criteria for synthetic peptides treat any specified impurity above it as disqualifying rather than as a trend to monitor. Sobre
A neutral example of how this data is generated: MOL Changes offers peptide synthesis at defined scale in classified cleanroom space, with analytical methods Peptídeos Sintéticos covering sequence confirmation and impurity profiling, so lot results arrive as a documented package the sponsor can drop into the CMC section.
Comprimidos 5: Documentação - Construindo a Narrativa IND CMC e o Conjunto de Perguntas Pré-IND
Documentation is where the other four pillars become reviewable. FDA IND CMC information requirements set out five items the drug substance section must contain: a description of the drug substance, including its physical, químico, and biological characteristics and evidence of structure; the name and address of the manufacturer; a general method of preparation; the limits and analytical methods used to assure identity, força, qualidade, e pureza; and stability data supporting the dosage form. Treat that list as the spine of a peptide CMC documentation checklist, then map each pillar onto it: sequence confirmation supplies the structure evidence, impurity profiling supplies the limits, analytical qualification supplies the methods, and process reproducibility supplies the lot-to-lot narrative behind the general method of preparation.
FDA frames the exercise as one of Phase 1 CMC flexibilities, describing the minimum information sponsors should submit and what can generally follow later. The page states that framing without enumerating which activities qualify, so the deferral list has to come from the linked guidance and be documented item by item.
Data integrity holds the narrative together. MHRA’s GxP data integrity guidance and definitions sets out the ALCOA principles (atribuível, legível, contemporâneo, original, preciso) that reviewers expect raw analytical records to satisfy.
Bring a written question set to the pre-IND meeting: which specifications can remain interim, what stability duration is acceptable at filing, whether qualification-level method data suffices, and how much detail the general method of preparation needs. Answers become the documented rationale.
⚠️ Aviso: A package that answers all five items but never states which activities are deferred, or why, is the failure mode. Reviewers read silence as an omission, not as flexibility.
Erros comuns que transformam um pacote pré-IND em retenção
The recurring failure in peptide CMC pre-IND packages is treating a reference framework as a requirement. Five mistakes account for most of the avoidable friction.
Quoting ICH Q3A thresholds as if they bound a peptide program. ICH Q3A was written for small-molecule drug substances, and its reporting, identification and qualification thresholds do not automatically govern a synthetic peptide. The mistake happens because the numbers are easy to find and easy to defend in an internal memo. The fix is to state which framework you are applying, por que, and what your own peptide impurity profiling data show at each threshold, rather than presenting a borrowed limit as settled.
Attributing requirements to FDA documents that do not contain them. Teams routinely cite “amino acid sequence” and “peptide map” as though FDA’s IND CMC page lists them. Não. Check what FDA’s IND CMC page does and does not say before a requirement enters your package, and label anything drawn from ICH, USP or internal policy as such.
Printing impurity numbers without a traceable source. Unverified figures for the ICH Q3A reporting and identification thresholds circulate widely in secondary summaries. If you cannot open the primary document and read the number, drop it or describe the threshold qualitatively.
Treating FDA’s Phase 1 CMC flexibilities as an enumerated deferral list. That page describes a flexible, phase-appropriate approach; it does not publish a checklist of what you may postpone. Build your own deferral rationale and document it.
Relying on a single-source echo cluster. Several trade write-ups of EndoCyclic’s March 2026 IND clearance announcement repeat one another without adding primary detail. Repetition is not corroboration. Go to the announcement itself and treat everything beyond it as unconfirmed.
⚠️ Aviso: The pattern behind all five mistakes is the same. A reference framework gets quoted as a requirement, and the package inherits a standard the program was never obligated to meet.
Como é o sucesso na reunião pré-IND
If the five pillars are locked and every deferral is documented with a rationale, the package you bring to the pre-IND meeting is one FDA can assess against the phase-appropriate standard rather than one that invites a CMC deficiency letter. Five outcomes make that concrete: a sequence traceable to raw data, an impurity profile with scope-stated limits, qualified methods with suitability data attached, a three-lot comparability overlay, and a narrative that maps onto the five drug substance items FDA expects.
The upside of arriving that way is measurable, with a caveat worth stating plainly. FDA estimates that sponsors of first-in-human Phase 1 INDs can reduce application development time by up to 12 months through FDA’s Phase 1 CMC flexibilities, the phase-appropriate minimum CMC information approach. FDA authors that policy and the estimate is not independent, so treat it as the agency’s own framing of the incentive rather than a benchmark from outside.
The direction of travel supports the same reading. FDA published o 17 peptide guidances FDA revised in July 2026 in a single day, covering impurity thresholds, higher order structure, atividade biológica, teste de resposta imune inata, and recombinant, sintético, and semi-synthetic peptide ANDAs. The agency is actively tightening its peptide vocabulary, which makes a package written in that vocabulary easier to review.
As a stretch goal, extend the comparability narrative you built for the pre-IND toward later-phase validation planning: the same three-lot overlay becomes the backbone of your Phase 2 and Phase 3 method validation strategy, so the work compounds instead of being redone.
Perguntas frequentes
A autorização ENDO-205 IND me diz qual pacote CMC o FDA aceitou?
Não. Marcha da EndoCyclic 2026 IND clearance announcement discloses the clearance, the indication, the preclinical basis and the planned Phase 1 design, and no CMC, fabricação, CDMO or synthesis detail. What the clearance does signal is narrower but still useful: a phase-appropriate package for a synthetic peptide was reviewable as submitted. It tells you nothing about which analytical methods, impurity limits or lot-data format the agency saw.
Os limites de impureza ICH Q3A se aplicam a substâncias medicamentosas peptídicas?
Não. ICH Q3A’s scope exclusion for peptides lists peptide among the drug substances the impurities guidance does not cover. Q3A remains a useful reference framework for structuring a peptide impurity profiling discussion, but it is not the binding threshold set for your drug substance. Peptide impurity control is risk-based, tied to the synthetic route, the deletion and truncation sequences the route can produce, and the toxicological qualification of each specified impurity. Produção de Peptídeos
O que o 0.5% o número especificado de impurezas relacionadas ao peptídeo realmente cobre?
It is a specified peptide-related impurity limit reported from FDA’s acceptance criteria for synthetic peptides. Two scope limits matter. It applies to specified peptide-related impurities, not to unspecified impurities, residual solvents or elemental impurities, which follow their own frameworks. And it should not be generalized to every peptide context. Verify the figure against the primary guidance document before you cite it in a pre-IND package.
Os métodos analíticos precisam ser totalmente validados antes da reunião pré-IND?
Não. FDA’s Phase 1 CMC flexibilities support phase-appropriate qualification at this stage, not full validation. What the meeting package does need is FDA’s IND CMC information requirements item on acceptable limits and analytical methods: you must show that each method is qualified for its intended use, that its limits of detection and quantitation are adequate for the impurity thresholds you are claiming, and that you can describe the qualification work you have done. Full validation belongs later, before the methods support release testing for clinical supply.
E se a confirmação da sequência falhar no primeiro lote?
Treat it as a route problem, not a documentation problem. Sequence confirmation failure on a first lot usually points to a coupling, deprotection or purification step that is not under control, and the fix is process work rather than a rewritten certificate of analysis. Re-run the confirmation on a fresh lot after you have identified the step, and document both the failure and the corrective action. A pre-IND package that shows you found and closed the gap is stronger than one that reports only passing lots.
Próximas etapas
You now have a gated, phase-appropriate peptide CMC documentation checklist across the five pillars: confirmação de sequência, perfil de impurezas, qualificação de método analítico, reprodutibilidade do processo, and the IND narrative itself. The standard to hold yourself to is the phase-appropriate minimum, and FDA’s own estimate of up to 12 months in reduced application development time is the reason it exists (FDA, recuperado 2026-06-09). Note the scope: FDA authors that policy, so the estimate is not independent. Once the three-lot comparability overlay is in place, the natural stretch goal is to extend it toward later-phase validation planning.
Principal vantagem: Five pillars, five verifiable artifacts: a sequence traceable to raw method output, an impurity profile with scope-stated limits, qualified methods with suitability data, a three-lot comparability overlay, and a narrative mapped onto FDA’s five drug substance items. If any one of those is missing, that is the gap to close before the pre-IND meeting.
Bring the package, not the questions. MOL Changes generates and documents lot-level analytical data for peptide drug substance, including sequence confirmation, perfil de impurezas, qualified methods, and lot-to-lot comparability, with synthesis from milligrams to kilograms in Class 100 cleanroom production and HPLC, EM, e testes de esterilidade. Talk to an expert, or request the lot-data package.
MOL Changes publishes as a peptide vendor and has a commercial interest in peptide quality standards. This article is not regulatory advice; sponsors should confirm requirements with FDA and consult qualified regulatory professionals.