क्या आईएसओ 9001:2026 पेप्टाइड लैब्स के लिए वास्तव में परिवर्तन
सेवाएं आईएसओ 9001:2026 पेप्टाइड प्रयोगशालाओं के लिए गुणवत्ता संबंधी निर्णय लेने और रिकॉर्ड करने के तरीके में परिवर्तन होता है, न कि कौन से रूप मौजूद हैं. पर संशोधन प्रकाशित किया गया था 16 सितम्बर 2026, ए पर योजना बनाई गई 36-आईएसओ/टीसी द्वारा समन्वित माह पुनरीक्षण चक्र 176 SC2 WG29, और यह आईएसओ के ग्यारह साल बाद उतरा 9001:2015. The खंड-दर-खंड परिवर्तन गिनती अभिलेख 6 बड़े बदलाव, 13 मध्यम, और 21 छोटा या संपादकीय.
जो खंड आगे बढ़े वे कुल से अधिक मायने रखते हैं. The मानक निकाय का खंड-दर-खंड सारांश कि संशोधन में क्या परिवर्तन होता है प्राथमिक संस्कृति और नैतिकता में परिवर्तन लाता है 5.1.1 और 7.3, और जोखिम को अवसर से अलग करता है 6.1.2 और 6.1.3. संशोधन के परिवर्तन-प्रबंधन खंड, 6.3 और 8.5.6, परिवर्तन के प्रबंधन के लिए अधिक एकीकृत दृष्टिकोण पेश करें. The 2024 जलवायु संशोधन को मूल पाठ में शामिल किया गया है 4.1.
किसी भी अभ्यास से पहले दो सीमाएँ बताने लायक हैं. आईएसओ 9001 पेप्टाइड एपीआई के लिए जीएमपी से नीचे बैठता है: यह एक सामान्य QMS ढाँचा है और इसे प्रतिस्थापित नहीं करता है आईसीएच गुणवत्ता दिशानिर्देश सेट जो एपीआई निर्माण को नियंत्रित करता है, ICH Q7 और ICH Q11. यह ISO/IEC भी नहीं है 17025 या आईएसओ 13485.
ट्रांज़िशन विंडो चलने की उम्मीद है 36 महीने, सितंबर के करीब प्रमाणपत्र की समाप्ति तिथि डाल दी गई है 2029, लेकिन वह तिथि अभी भी लंबित है IAF संकल्प और प्रमाणन-निकाय के सूत्र असहमत हैं. IAF संकल्प के विरुद्ध इसकी पुष्टि करें, विक्रेता पृष्ठ नहीं.
क्यों 2026 पुनरीक्षण एक सिस्टम प्रश्न है, कागजी कार्रवाई का सवाल नहीं
संशोधन साक्ष्य की भाषा को फिर से लिखता है, अभिलेखों की सूची नहीं. आईएसओ 9001:2026 "दस्तावेज़ जानकारी बनाए रखें" और "दस्तावेज़ जानकारी बनाए रखें" के पक्ष में "दस्तावेज़ जानकारी के रूप में उपलब्ध होगी" और "दस्तावेज़ जानकारी साक्ष्य के रूप में उपलब्ध होगी" को हटा दिया गया है।," और यह जहां रिकॉर्ड मौजूद होना चाहिए उसकी खंड-दर-खंड सूची अनुबंध A.2 e के माध्यम से पुष्टि करता है) वह "साक्ष्य के रूप में" कोई कानूनी साक्ष्य मानक नहीं बनाता है.
गिनती अपरिवर्तित है. खंडों के पार 4 को 10, 27 बिंदुओं के लिए दस्तावेजी जानकारी की आवश्यकता होती है: 8 दस्तावेज़ के रूप में उपयोग के लिए उपलब्ध है, 19 अभिलेखों के रूप में साक्ष्य के रूप में उपलब्ध है, जैसा ही सेट है 2015. इसलिए एक कस्टम पेप्टाइड संश्लेषण गुणवत्ता प्रणाली को अधिक कागजी कार्रवाई की आवश्यकता नहीं होती है. इसे ऐसे निर्णयों की आवश्यकता है जिनका पता बैच से लगाया जा सके.
कदम 1: दस्तावेज़ीकरण को छूने से पहले मैप पेप्टाइड बैच ट्रैसेबिलिटी
बैच रिकॉर्ड से प्रारंभ करें, गुणवत्ता मैनुअल नहीं. पेप्टाइड बैच ट्रैसेबिलिटी रीढ़ की हड्डी है जो हर बाद के अभ्यास पर टिकी रहती है: नियंत्रण बदलें, जोखिम आकलन, आपूर्तिकर्ता निरीक्षण, और ग्राहक दस्तावेज़ीकरण सभी बहुत कुछ से जुड़े होते हैं जिनका कच्चे माल से जारी होने तक अनुसरण किया जा सकता है.
कारण संरचनात्मक है. ठोस-चरण पेप्टाइड संश्लेषण (एसपीएसएस) एक समय में एक अमीनो एसिड एक राल पर एक श्रृंखला बनाता है, और एक विशिष्ट 30-अमीनो-एसिड पेप्टाइड की आवश्यकता होती है 30 अनुक्रमिक युग्मन और संरक्षण चक्र. पर 99% प्रति चरण दक्षता, संचयी उपज हानि और अशुद्धता संचय महत्वपूर्ण हो जाता है, जैसा न्यूलैंड लैब्स ने अपने विवरण में बताया है कि पेप्टाइड प्रक्रिया के बढ़ने पर क्या परिवर्तन होते हैं. एक एकल 30-मेर भी उत्पन्न कर सकता है 50 को 100 संभावित अशुद्धियाँ, और न्यूलैंड लैब्स बताते हैं कि पेप्टाइड अशुद्धियों को अपनी स्वयं की नियंत्रण रणनीति की आवश्यकता क्यों है: पर या उससे ऊपर की अशुद्धियाँ 0.10% संरचनात्मक पहचान की आवश्यकता है, और ऊपर वाले 0.5% सुरक्षा औचित्य की आवश्यकता है.
यही कारण है कि अंकगणित एक राल बहुत कुछ है, एक अमीनो एसिड लॉट, या मध्य-अभियान कॉलम स्वैप को उस सटीक पेप्टाइड लॉट के लिए ट्रेस किया जाना चाहिए जिसे उसने छुआ था. पहले वंशावली बनाओ: कच्चे माल के लॉट के लिए पेप्टाइड लॉट, प्रक्रिया पैरामीटर, प्रक्रियाधीन जाँचें, विश्लेषणात्मक परिणाम, विचलन, और रिलीज़ रिकॉर्ड. फिर इसे सत्यापित करें. एक जारी किए गए लॉट को चुनें और रिकॉर्ड सेट को छोड़े बिना उसके कच्चे माल के लॉट का पीछे से पता लगाएं. यदि आपको किसी लिंक को बंद करने के लिए किसी को फोन करना है, नक्शा ख़त्म नहीं हुआ है.
कदम 2: विश्लेषणात्मक विधि संस्करणों और कॉलम स्वैप के आसपास परिवर्तन नियंत्रण बनाएं
अभियान के मध्य में कॉलम स्वैप एक नियोजित परिवर्तन है, लैब हाउसकीपिंग नहीं. अंतर्गत आईएसओ 9001:2026 खंड 6.3, मानक अब जिस नियोजन इनपुट से बदलाव की अपेक्षा करता है वह उद्देश्य और संभावित परिणाम हैं, क्यूएमएस की अखंडता, संसाधनों और सूचना की उपलब्धता, जिम्मेदारियों और प्राधिकारियों का आवंटन या पुनर्आवंटन, परिवर्तन का संचार कैसे किया जाता है, प्रभावशीलता की निगरानी कैसे की जाती है, और परिणामों की समीक्षा कैसे की जाती है. कॉलम को अनपैक करने से पहले उन्हें परिवर्तन प्रपत्र में लिखें.
कारण विश्लेषणात्मक है, प्रशासनिक नहीं. लक्ष्य पेप्टाइड के साथ विलोपन अनुक्रम और एपिमर्स लगभग आइसोबैरिक हैं, इसलिए रिवर्स-चरण एचपीएलसी पृथक्करण ऑर्थोगोनल तरीकों के बिना अविश्वसनीय है, और न्यूलैंड लैब्स सह-संशोधन को अपवाद के बजाय आदर्श के रूप में वर्णित करता है. एक नया कॉलम लॉट या ग्रेडिएंट अवधारण और रिज़ॉल्यूशन को बदल देता है, यही कारण है कि योजना बनाने के लिए अपवाद के बजाय सह-संयोजन आदर्श है. इसलिए पेप्टाइड संश्लेषण में परिवर्तन नियंत्रण को विधि संस्करण को कवर करना होगा, स्तंभ स्थल, और एक नियंत्रित सेट के रूप में ग्रेडिएंट, एक परिभाषित प्रभावशीलता जांच के साथ: बनाए गए संदर्भ नमूने को दोबारा चलाएं और नया डेटा जारी करने से पहले ऐतिहासिक परिणाम से तुलना करें.
जांच सूची: खंड 6.3 परिवर्तन-योजना इनपुट
परिवर्तन का उद्देश्य और इसके संभावित परिणाम
पेप्टाइड संश्लेषण Effect on the integrity of the quality management system
Availability of resources and of information
Allocation or reallocation of responsibilities and authorities
How the change is communicated
How effectiveness is monitored and how results are reviewed
कदम 3: संश्लेषण और प्रक्रिया-नियंत्रण में जोखिम-आधारित सोच लागू करें
आईएसओ 9001:2026 splits clause 6.1 into two subclauses: risk management (6.1.2) and opportunity-based thinking (6.1.3). For a peptide lab, that separation forces something most quality manuals have never done: name the specific failure modes of solid-phase synthesis, not the generic categories.
Build the register from chemistry. The practitioner primer on what goes wrong in solid-phase synthesis documents incomplete couplings that leave truncated and deletion sequences, aspartimide formation at Asp that drives epimerization and a −16 Da mass shift, and C-terminal Cys on Wang resin eliminating to dehydroalanine (−36). Each entry needs a control and a detection point, not just a severity score.
Scale changes the risk profile. A CDMO’s account of what changes when a peptide process scales identifies mass transfer (राल की सूजन, solvent exchange, mixing dynamics), heat transfer (तापमान दुकान gradients altering epimerization rates), and downstream loadability (reduced preparative column resolution forcing pooling changes) as the variables that shift between mg and kg.
कुंजी ले जाएं: No single ICH framework governs peptide impurity thresholds. ICH Q3A and Q3B explicitly exclude synthetic peptides, so your register has to justify its own action limits.
कदम 4: पेप्टाइड आपूर्तिकर्ताओं को गंभीरता के आधार पर योग्य बनाएं, सर्टिफिकेट से नहीं
Supplier qualification should follow the consequence chain, not the certificate portfolio. A resin lot change mid-campaign shifts swelling behaviour and coupling kinetics, an amino acid lot with different enantiomeric purity shifts the epimer profile, and a solvent lot change alters the ion-pairing behaviour that governs how much residual TFA remains in the crude. That last consequence is not cosmetic: residual TFA from solid-phase synthesis is considered undesirable or potentially toxic in clinical use, and removing it by counterion exchange to chloride or acetate adds process steps (न्यूलैंड लैब्स, पुनर्प्राप्त 2026-04-27).
So qualification depth should track what the material does to your process. Materials that can move a critical quality attribute warrant certificate review plus independent analytical verification; materials that can halt a campaign warrant an audit. And a certificate does not by itself prove an operating system: it evidences one lot against one specification, not the change control, विचलन प्रबंधन, and investigation records behind it. Ask for those records instead of the certificate alone.
कदम 5: संस्करण नियंत्रण और विचलन समीक्षा से बचे रहने वाले क्लाइंट दस्तावेज़ीकरण जारी करें
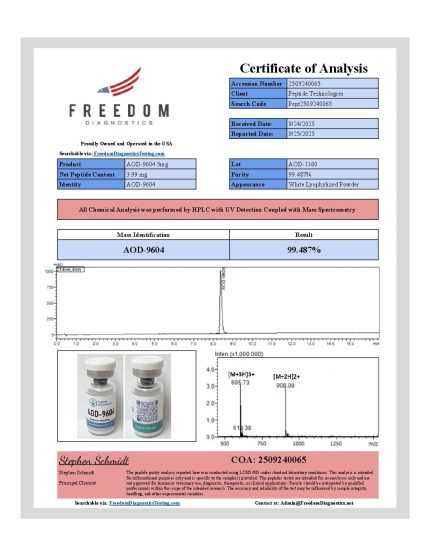
इस चरण के अंत तक, every client-facing document you issue can be traced back to the exact conditions that produced it. That traceability is what lets a package survive a deviation review months later.
The clause-by-clause list of where records must exist separates two things that peptide labs often blur together. Documents are controlled, मौजूदा, approved, and available where the work is done; records are protected from alteration, retained for a defined period, and retrievable on demand (आईएसओ 9001:2026 documented information listing, पुनर्प्राप्त 2026-09-18). A certificate of analysis is a record. The method it was run under is a document.
That distinction has teeth in a custom peptide synthesis quality system. A CoA, an HPLC chromatogram, and a mass spectrum (एमएस, which measures a molecule’s mass-to-charge ratio to confirm identity) issued per lot must each point to the method version and column lot in force on the day of analysis. Change the column mid-campaign and the earlier chromatograms still belong to the earlier method version. If the package does not say so, a later deviation review cannot reconstruct which conditions applied to which lot.
MOL Changes operates an integrated peptide synthesis platform with Class 100 ultra-sterile cleanroom production and mg-to-kg scale-up, issuing a CoA, HPLC chromatogram, and MS spectrum per lot. That is a documentation capability, not a certification claim.
बचने के लिए सामान्य गलतियाँ
Treating the revision as a documentation reformatting exercise. The clause-by-clause list of where records must exist is unchanged at 27 points from 2015, so teams reasonably conclude that existing templates will pass. What actually moved is how decisions सिंथेटिक पेप्टाइड्स get recorded and linked: risk reasoning, change justification, and the traceability between them. The fix is to audit your decision trail, not your document set. Ask whether a reviewer could reconstruct why a specification changed, not whether a form exists.
Qualifying suppliers by certificate rather than by criticality. A supplier certificate says a system was audited; it says nothing about the resin and amino acid lots that drive epimerization and truncation. Rank suppliers by the failure modes their materials can cause, then set oversight depth accordingly.
Running change control on synthesis steps only. Analytical method versions and column swaps sit outside many change processes, which is where co-elution risk enters. Co-elution is the norm rather than the exception in peptide analysis, so a column change can silently alter what a release result means.
Building a risk register from generic categories. A register organized around “supplier risk” and “process risk” satisfies an auditor and informs no one. Name the peptide failure modes you are actually managing.
परिणाम: कार्य प्रणाली कैसी दिखती है
A working ISO 9001:2026 for peptide labs system is recognizable by what you can retrieve, not by what you can display. Five markers are checkable without an auditor in the room. के बारे में
A released lot traces backwards to its raw-material lots without leaving the record set. Every analytical method version and column lot in force on an analysis date is recoverable. The risk register names peptide-specific failure modes, each with a control and a detection point. Supplier qualification depth varies by material criticality rather than by certificate count. And a client package issued eighteen months ago still reconciles against the method version and deviation log of its day.
The stretch goal is extending that same genealogy spine to cold-chain excursions and to endotoxin and particulate risk for any lot destined for clinical work. पेप्टाइड उत्पादन
This is an operating-model explainer, not regulatory advice. एमओएल चेंजेस की पेप्टाइड गुणवत्ता मानकों में व्यावसायिक रुचि है; its documented QC stack, per-lot CoA and HPLC/MS documentation, और कक्षा 100 cleanroom are capability facts, not certification claims.
अक्सर पूछे जाने वाले प्रश्नों
आईएसओ है 9001:2026 पेप्टाइड प्रयोगशाला के लिए प्रमाणीकरण आवश्यक है?
No regulator requires it. आईएसओ 9001 is a voluntary standard, and certification is a customer-driven expectation rather than a legal obligation for most peptide laboratories. What is mandatory depends on what you make and where you sell it: a peptide intended as an active pharmaceutical ingredient falls under GMP, और यह आईसीएच गुणवत्ता दिशानिर्देश सेट जो एपीआई निर्माण को नियंत्रित करता है takes over from there. आईएसओ 9001 and GMP are not substitutes. A quality management system built to ISO 9001 can sit underneath a GMP framework, but it does not discharge GMP obligations on its own.
36 महीने की संक्रमण अवधि कैसे काम करती है?
The certification body’s revision timeline gives certified organizations a 36-month window to move from the 2015 edition to the 2026 revision. Certificates issued against the older edition remain valid during that window, so there is no immediate deadline pressure. The practical risk sits elsewhere: audits scheduled early in the transition will expect to see evidence that your gap analysis is underway, even if the full transition is not complete. Start the mapping now, not in month 30.
दस्तावेज़-सूचना सेट वास्तव में कितना बदल गया?
Less than the revision headlines suggest. The clause-by-clause list of where records must exist shows 27 points unchanged between editions. The shift is in what the standard asks you to do with those records, particularly around risk, नियंत्रण बदलें, and organizational knowledge, not in the number of documents you must hold. If your 2015 system is well built, you are extending it rather than rebuilding it.
एक छोटे प्रयोगशाला क्रम को सबसे पहले क्या करना चाहिए??
Start with batch traceability. It is the practice every other one depends on: नियंत्रण बदलें, risk-based in-process decisions, and client documentation all assume you can reconstruct what happened to a lot. A lab that can trace a batch end to end has the foundation to layer the remaining four practices on top. A lab that cannot will find each of them harder to implement than it needs to be.
निष्कर्ष
The five milestones are one system, not five projects. Batch traceability gives change control something to point at; change control gives risk-based thinking a place to land; supplier qualification extends the same logic upstream; and client documentation is simply the visible surface of all four. Break any link and the chain fails at the point a reviewer tests it.
That is the difference between a certificate on a wall and a quality system that holds up. A certificate records that an auditor once found your documentation in order. A working system survives the harder test: a deviation review six months into a campaign, or a method change that forces you to show which lots were released under which version. आईएसओ 9001:2026 for peptide labs rewards the second, because its documented-information expectations follow the product through synthesis, विश्लेषण, and release rather than stopping at the binder.
If you take one action from this guide, review your own evidence trail against the 27 documented-information points and see where the chain breaks first. If you would rather see what a complete per-lot package looks like, MOL Changes publishes an example of the CoA, एचपीएलसी, and MS documentation it issues per lot.