전제조건: ISO를 매핑하기 전에 팀에 필요한 것 9001:2026 펩타이드 R에&디
This is a translation exercise, 인증 감사가 아님. 그만큼 2026 edition updates the framework rather than replacing it, so a team already running ISO 9001:2015 can adopt the changes without redesigning its system (ANSI’s release announcement, 16 구월 2026). Budget one working session per artifact, not a re-implementation project.
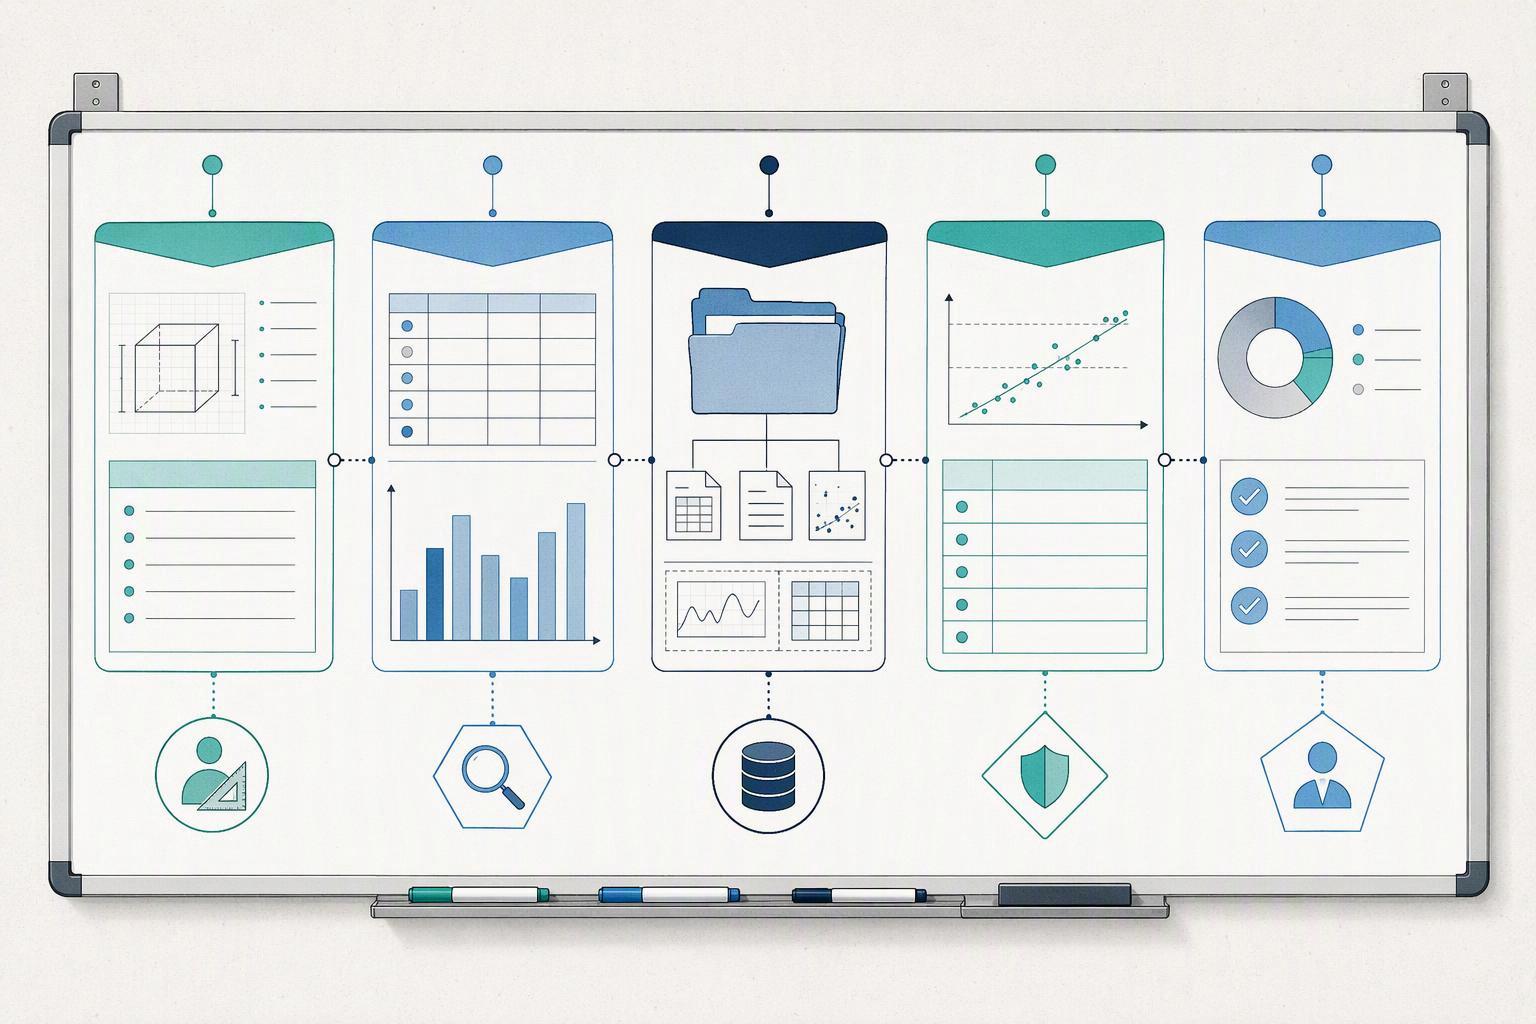
Before you start, confirm you can open five controlled documents: the current specification sheet, the deviation log, the raw-data archive, the CoA template, and the release or quarantine decision record. Name the owner of each: specification approver, change-control owner, deviation signatory, raw-data reviewer, release decision-maker. Assume the team is comfortable with RP-HPLC purity reporting, counterion forms and lot-to-lot comparison.
The revision strengthens requirements for quality culture and ethical behaviour, with leadership explicitly responsible for the environment, and it separates risks from opportunities (BSI’s key-changes guidance, 2026). That is why the checklist covers artifacts and named roles rather than documents alone.
⚠️ 경고: This article describes quality-system practice, not regulatory or medical advice, and does not interpret the standard on a certification body’s behalf.
단계 1: Translate Quality Culture and Leadership Into Named Accountabilities
The first thing ISO 9001:2026 asks of a peptide R&D group is not a document but an assignment. Quality culture, the shared set of values and behaviours that determines whether people follow the system or work around it, moves from implied to explicit in the revision, and leadership is named as the party responsible for creating the environment in which it holds (ANSI, 2025 change summary). Ethical behaviour is strengthened alongside it.
That clause becomes actionable only when each accountability has a name, an artifact and a signature. A peptide quality management system fails at this point more often than at any technical one: the manual says the quality unit approves specifications, but no signature block records who did, so a deviation closes without an owner.
|
Accountability |
Artifact that evidences it |
Role that signs |
|---|---|---|
|
Specification approval |
Controlled specification sheet, versioned |
Quality unit head |
|
변경 제어 |
Change request with impact assessment |
Process owner |
|
펩타이드 1 Deviation closure |
Investigation report with root cause |
QA signatory |
|
Raw-data review |
Biosynthesis Peptide Reviewed chromatogram and mass-balance package |
Analytical reviewer |
|
Release or 펜타펩타이드 4 quarantine |
Batch disposition record |
Authorised release Peptide Biosynthesis officer |
Counterion and salt-form variability, caught by ion chromatography, conductivity and mass-balance checks, is a useful test case: if nobody is named as the analytical reviewer for those checks, the accountability exists on paper only (Merck, 검색됨 2026-02-27).
단계 2: Build Controlled Specifications That Survive a Lot-to-Lot Comparison
A peptide specification is only controlled when it states the sequence, the identity method, the purity limit and the method behind it, the salt or counterion form, and the limits for residual counterions, residual solvents and water content. 그만큼 합성 펩타이드의 개발 및 제조에 관한 EMA 가이드라인 defines exactly these parameters, adding sterility and endotoxin limits where the intended use requires them, plus storage and shipping conditions.
Counterion variability is where lot-to-lot comparison usually breaks down. The same sequence supplied as the trifluoroacetate salt in one lot and the acetate salt in the next shows a different apparent mass, different solubility and different chromatographic behaviour, which is why ion chromatography, conductivity and mass-balance checks belong in the release decision alongside the purity figure.
The standard leaves room for interpretation here: it requires controlled specifications without prescribing which counterion limits apply to a given peptide, so your quality management system has to state that choice, version it, and defend it in the specification sheet rather than in a reviewer’s memory.
단계 3: Turn Deviations Into Learning With Two-Phase Investigation and Trending
A deviation is closed when the investigation has produced a cause, a corrective and preventive action (CAPA), and verified effectiveness, not when the batch is dispositioned. 그만큼 FDA guidance on investigating out-of-specification test results sets a two-phase sequence: 단계 1 stays in the laboratory, covering raw data, instrument status, sample preparation, analyst technique and method performance. Only if no assignable lab cause is found does Phase 2 move to manufacturing, covering the synthesis and batch record, 프로세스 단계, sampling representativeness, equipment and related batches. Root-cause tools include 5 Whys, fishbone diagrams, FMEA and fault-tree analysis, and CAPA needs effectiveness verification before closure.
Enforcement data shows where this breaks down. Complere’s tabulation of FDA’s FY2025 cited-regulation spreadsheet 기록 243 citations under §211.22(디) for quality-unit procedures and 164 under §211.192 for failure to investigate discrepancies, ~에 맞서 14, 10 그리고 9 for incomplete laboratory records, 가로질러 713 drug 483s from 1 십월 2024 에게 30 구월 2025.
팁의 경우: Those FY2025 counts are a floor, not a census. FDA’s spreadsheet covers system-generated 483s only, so manually prepared forms are excluded.
For peptide specification and deviation control, the move that turns records into learning is trending by synthesis step, purification step, assay method, operator and supplier rather than by exception count. 집합, incomplete deprotection and counterion drift then surface as patterns instead of isolated events.
단계 4: Make Data Integrity Real With a Reconstructable Raw-Data Package
Data integrity in peptide R&D is not a policy statement. It is whether a reviewer can reconstruct the run from what you archived. Data validation, in this context, means demonstrating that the record you keep is complete, accurate and traceable back to the instrument that produced it.
A defensible raw-data package carries the native original data plus the metadata needed to rebuild the analysis: full chromatograms, integration and processing methods, system and method parameters, analyst identity, software and firmware versions, calibration and qualification state, and the audit trail showing every change, deletion and the reason for it (PIC/S Guidance on Data Integrity, PI 041-1, 2021). 그만큼 MHRA GxP Data Integrity Guidance and Definitions (2018) adds the test that matters in an audit: the record must be capable of reconstructing the activity and associating every change to a person, a date and time, and a reason for change.
Two obligations are easy to conflate. USP <1058> governs instrument qualification, establishing that the instrument is fit for use before a method runs on it. The ICH Q2(R2) guideline on validation of analytical procedures (2023) governs the procedure lifecycle instead, and requires the validation protocol to be written before the study, with the reportable range confirmed by acceptable response, accuracy and precision. Instrument fitness and method validity are separate claims, and a peptide Certificate of Analysis documentation package that shows only one of them is incomplete.
실패 모드는 익숙하다: a purity figure on a CoA while the chromatogram, integration settings and calibration state sit in a different folder, or nowhere. 아래에 21 CFR 부분 11 and CGMP laboratory-record expectations, electronic records captured electronically must be retained in electronic form, and the laboratory record must hold a complete record of all data secured during testing, including graphs, charts and spectra (FDA, 날짜가 기재되지 않은).
단계 5: Report Transparently So a CoA Says What It Actually Certifies
A Certificate of Analysis is evidence about one lot, tested by one method on one date. It certifies the measured result for that identified batch against the supplier’s stated release specification, and it does not certify other lots, future shipments, storage stability, untested attributes, or the supplier’s compliance for your intended use (Peptigraph; CASRAI).
That boundary is where most peptide Certificate of Analysis documentation breaks down. A numeric impurity result is only comparable across reports if the method version is stated, and a “not detected” result is only interpretable if the method’s limit of detection or quantitation is disclosed, because absence of detection is not absence of analyte (Peptigraph).
핵심 내용: A “not detected” result is only interpretable when the method’s LOD/LOQ and method version are disclosed. Absence of detection is not absence of analyte.
Five disclosure fields carry that transparency: method version, LOD/LOQ, 반대이온 형태, the specification version the lot was released against, and the raw-data package reference. 그들 없이는, peptide data integrity ALCOA+ stops at the signature block. Peptide Bond Synthesis
실패 모드는 조용하다. A team compares a 0.3% impurity figure against a previous supplier’s report produced under a different method version, reads the difference as a trend, and acts on a comparison the two documents never supported.
단계 6: Close the Loop Across Client and Laboratory Teams
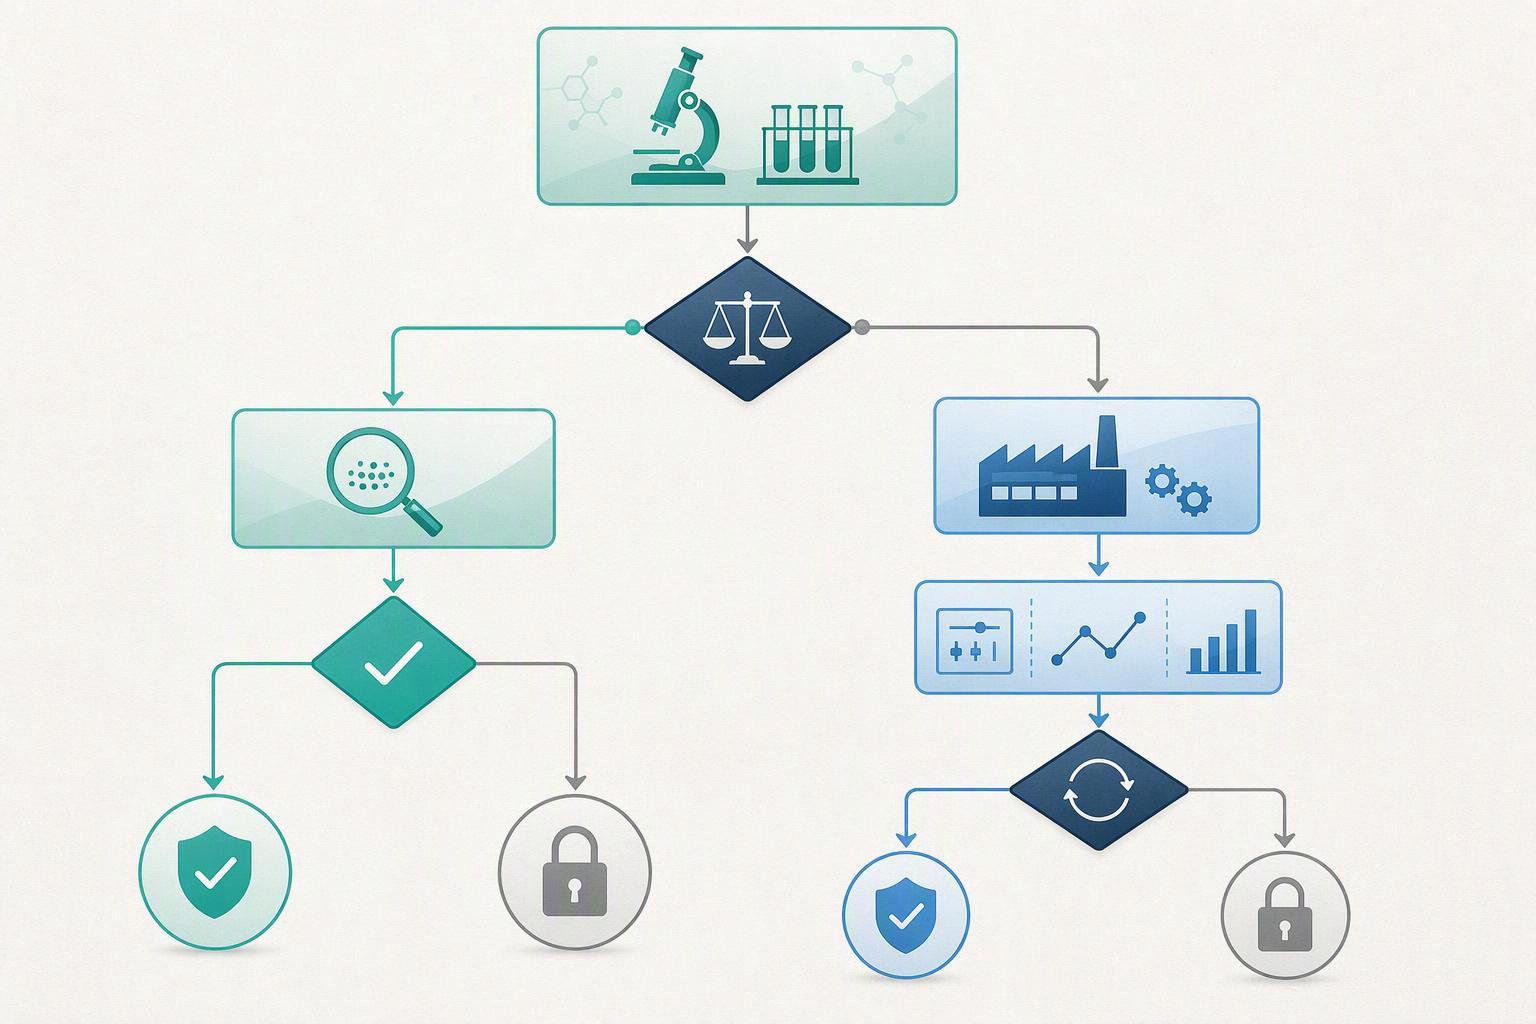
The accountability split only holds if both sides agree in writing on who owns the specification, who owns change control, who signs a deviation, who reviews raw data, and who makes the release or quarantine call. Revisit that agreement at tech transfer, not after the first failed lot. 그만큼 2026 revision gives greater attention to change management, which makes an undocumented split between client and laboratory a finding waiting to happen.
Peptide work shows why the split matters. On-resin aggregation appears as resin shrinkage and slowed coupling, and incomplete deprotection is confirmed chromatographically on the cleaved product. A partner able to supply lot-specific analytical documentation closes that loop faster.
팁의 경우: In-process color tests can return false negatives under severe aggregation. Confirm on the cleaved product before you treat a step as complete. 펩타이드
피해야 할 일반적인 실수
The most frequent failure is treating the revision as a documentation refresh: rewriting the quality manual while the specification sheet, the deviation log and the raw-data archive stay exactly as they were. Five mistakes account for most of the gap between a compliant-looking system and one that holds up.
Validating the method only at release. Teams confirm the batch meets specification but never confirm the analytical method still performs. 고치다: treat method validation as a gate before release, not a one-time event.
Closing deviations on disposition. A deviation is marked closed once the batch is released, with no check that the corrective action worked. 고치다: require verified CAPA effectiveness before closure.
Archiving processed results without native data. The report is filed; the instrument file and audit trail are not. 고치다: archive the reconstructable package, not the summary.
Issuing CoAs without method version or LOD/LOQ. A numeric result is only comparable across reports if the method version is stated. 고치다: put method version and detection limits on every CoA.
Leaving the client–laboratory accountability split undocumented. 고치다: name the owner for each record at tech transfer.
메모: The FY2025 data-integrity share of FDA warning letters comes from a vendor analysis of public enforcement data. Corroborate it against FDA’s own database before quoting it internally.
자주 묻는 질문
Does the 2026 revision require us to rebuild our quality management system?
아니요. 그만큼 2026 edition updates the framework rather than replacing it, and organizations already on the 2015 version can adopt it without redesigning their system, per AFNOR’s international transition FAQ. What changes is emphasis: quality culture, risk and opportunity, and data integrity become explicit expectations. Your documented processes stay; the evidence attached to them gets sharper.
What is the actual transition deadline for ISO 9001:2015 인증서?
ISO 9001:2015 certificates are valid only until the transition end date, AFNOR reports, with the three-year window closing on 30 구월 2029. The same guidance advises scheduling a 2026-based audit by April 2028, and no 2015-based audits are scheduled from May 2029. The rule originates with the accreditation community rather than ISO itself, so confirm dates with your certification body.
Do ISO/IEC 17025 or cGMP replace ISO 9001 for a peptide laboratory?
아니요, they cover different aspects. ISO/IEC 17025 addresses the competence of testing and calibration laboratories, so it speaks to whether a purity assay or impurity method is technically defensible. cGMP sets manufacturing standards for pharmaceutical production. ISO 9001 governs the quality management system that ties specifications, 편차, records and responsibilities together. When a client requests certification evidence, ask which of the three they mean.
What should we do when a supplier’s CoA reports “not detected” without an LOD?
Request the method’s limit of detection or quantitation and the method version before accepting the result. Absence of detection is not absence of analyte: “not detected” only means the substance fell below a threshold the CoA does not state, and that threshold may sit above the level your specification cares about. Without the LOD, method version and instrument conditions, treat the figure as unverified and re-test in-house or qualify the material against a method you control.
결론
You now have the six pieces that turn ISO 9001:2026 from a governance principle into daily peptide R&D practice: named accountabilities, version-controlled specifications, a two-phase deviation process with trending, a reconstructable raw-data package, a CoA that says what it certifies, and a written client–laboratory split. None of these require rebuilding your quality management system; they require making what already exists traceable.
The reason to start now is the calendar, not a metric. The transition window for ISO 9001:2026 closes on 30 구월 2029 (ISO, 검색됨 2026-06-11), which sounds distant until you count the audit cycles, supplier requalification and document revisions a peptide programme needs to get there.
다음 단계: Request the documentation package or a technical feasibility assessment to see how these practices map to your current ISO 9001:2026 peptide R&D workflow. It is a low-commitment first conversation, not a purchase.
폭로: MOL Changes는 펩타이드 품질 표준에 상업적인 관심을 갖고 있습니다..