Peptide Analytisch Gereedheidskader: Nitriet om vrij te geven
Inhoudsopgave
Wat is een Peptide Analytical Readiness Framework?
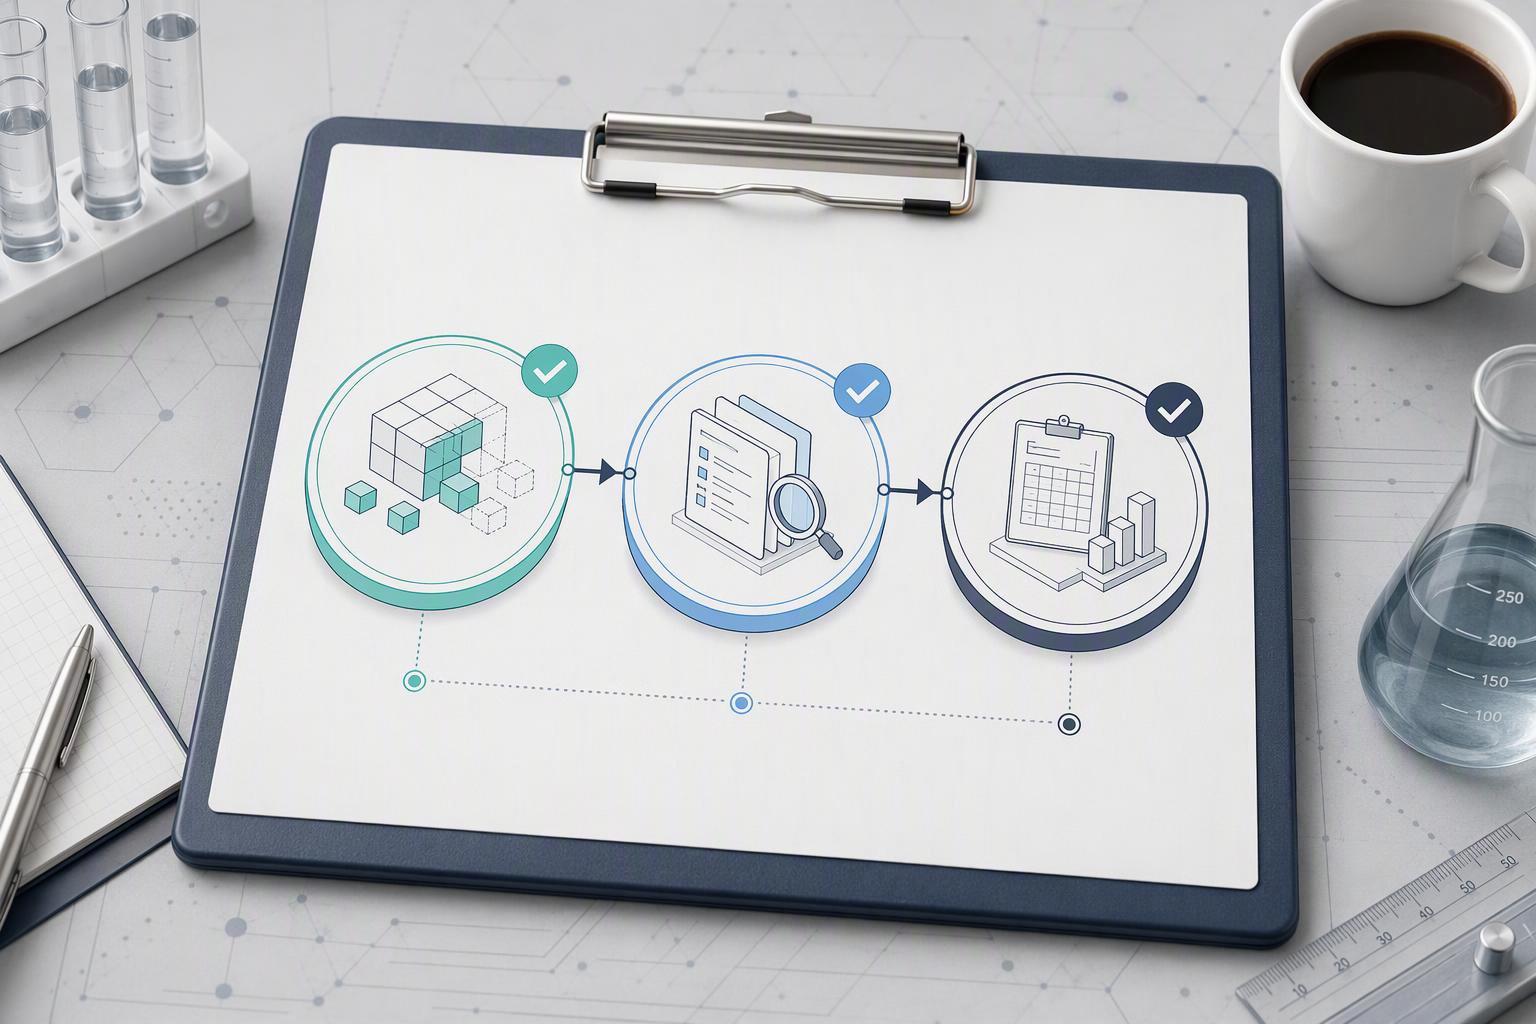
Een raamwerk voor analytische paraatheid voor peptiden is de gedefinieerde identiteitsset, zuiverheid, residu, degradatie, verpakkingsinterface en documentatiebewijs dat een CMC-team moet bewaren voordat de formulering wordt vergrendeld of klinische levering plaatsvindt. Het is een beslissingsstructuur, geen testlijst: elk domein beantwoordt een specifieke vraag die een recensent zal stellen, en de zes samen vertellen je of de medicijnsubstantie goed genoeg gekarakteriseerd is om verder te komen.
Peptiden voldoen niet helemaal aan de standaard checklist voor kleine moleculen. EMA’s richtlijn voor synthetische peptiden, herzien in 2022 en effectief vanaf 2023, behandelt synthetische peptiden als een aparte klasse, juist vanwege hun onzuiverheidsprofiel, afbraakroutes en analytische controlestrategieën verschillen van zowel kleine moleculen als biologische stoffen. De universele tests en identiteitscriteria van Q6A blijven de basis waaraan elke peptide-inzending nog steeds moet voldoen; de zes domeinen hier breiden die basislijn uit in plaats van deze te vervangen.
Dat is de praktische waarde van het raamwerk: Hierin staat welke checklist daadwerkelijk van toepassing is, en waar de generieke niet meer voldoende is.
Eén grens voordat we verder gaan. Dit raamwerk blijft op analytische en CMC-grond. Het gaat niet over klinische dosering, therapeutische selectie of patiëntgerichte beslissingen, en niets vervangt hier een strategie voor het indienen van regelgeving, beoordeeld door uw eigen kwaliteits- en regelgevende functies.
Waarom het nitrietsignaal een gereedheidstrigger is, Geen enkele test
Een nitrietresultaat is een risicomarkering, geen bewijs dat er zich een nitrosamine heeft gevormd. It tells you that a precursor pool may exist somewhere in your peptide’s route, and that the right response is a readiness review across identity, zuiverheid, residuen, degradatie, packaging and documentation, not a single confirmatory assay bolted onto the release panel.
The regulatory stakes explain why. FDA’s NDSRI acceptable-intake guidance sets a conservative acceptable intake of 26.5 ng/day for nitrosamine drug substance-related impurities, and FDA’s potency-category limit table assigns categories under the CPCA framework rather than one blanket number. EMA’s three-step nitrosamine framework expects marketing authorisation holders to run step 1 risk evaluation, step 2 confirmatory testing where warranted, and step 3 mitigation, with corrective actions typically expected within three years. De verwachting van Health Canada dat het per lot vrijkomt, voegt eraan toe dat sommige producten nitrosaminetests nodig hebben bij batchvrijgave in plaats van één keer bij indiening.
Bronnen zijn breder dan de API. In de richtlijnen voor nitrosaminecontrole van de FDA worden hulpstoffen genoemd, proceswater en bepaalde verpakkingsmaterialen als potentiële bijdragers, en noch de FDA, noch de EMA publiceert een procentuele verdeling waaruit blijkt welke bron domineert in peptideprocessen. Die afwezigheid is op zichzelf al het argument voor een raamwerk: je kunt bronnen niet uitsluiten of uitsluiten door aannames.
Eén eerlijke beperking hoort hier thuis. Omgekeerde-fase-HPLC met UV-detectie bij 214–220 nm, de zuiverheidsmethode van het werkpaard, kan lage of slecht gescheiden onzuiverheden missen, een schoon UV-chromatogram is dus geen bewijs dat het nitrietsignaal is opgelost.
Sleutel afhaalmaaltijd: De limieten voor aanvaardbare inname gelden per onzuiverheid en variëren per autoriteit, 26.5 van/dag onder FDA-categorie 1 tegen 18 van/dag onder de categorie Health Canada 1, een enkel getal mag dus nooit worden geciteerd zonder zijn autoriteit en reikwijdte.
Domein 1: Identiteit – Waarom één methode nooit genoeg is
Identiteitstests voor een synthetisch peptide kunnen niet op één enkele techniek berusten. De universele tests en identiteitscriteria van ICH Q6A vereisen een identificatietest die onderscheid maakt tussen nauw verwante structuren, en de dichtstbijzijnde structurele buren van een peptide zijn precies degenen die met één enkele methode moeilijk te scheiden zijn: een afknotting waarbij één residu ontbreekt, een deletie aan de N-terminus, of een sequentievariant die verschilt door een enkel aminozuur.
Orthogonale methoden werken omdat ze allemaal een andere eigenschap ondervragen. Massaspectrometrie bevestigt de moleculaire massa tegen de sequentie-afgeleide theoretische waarde; peptide mapping with LC-MS/MS locates where the sequence diverges; amino acid analysis confirms composition; capillary electrophoresis separates by charge-to-size ratio rather than hydrophobicity. Two methods built on independent principles give a defensible identity claim, because a co-eluting impurity that fools one separation principle is unlikely to fool a mass measurement as well.
The failure mode is a single-retention-time identity test that passes a truncated sequence because the deletion shifts retention only marginally. Pairing a separation method with a mass-based method closes that gap.
Domein 2: Zuiverheids- en onzuiverheidsprofilering — Instellen van fase-geschikte drempels
Peptide impurity profiling and release testing does not inherit the small-molecule threshold triad. EMA’s synthetic-peptide guideline sets peptide-specific reporting, identificatie Aangepaste gesynthetiseerde peptidenand qualification thresholds at 0.1%, 0.5% En 1.0% respectievelijk, met een 0.1% quantitation limit for the LC purity method. Those are the numbers to design against.
The thresholds readers often carry over come from elsewhere. Ik Q3A(R2) threshold table governs new drug substances, and ICH Q3B(R2) degradation-product thresholds apply to drug products, where the limits are keyed to maximum daily dose rather than fixed. A flat 0.1/0.2/1.0 set is not what Q3B says.
Forced degradation is how you show the method can carry those thresholds instead of asserting it. Stress the peptide, then demonstrate resolution of the resulting species at the quantitation limit.
De faalmodus is stil: a method that cannot resolve a co-eluting truncation sequence reports a passing purity. The number looks clean because the impurity was never separated from the main peak.
Voor Tip:Q3B thresholds are dose-keyed. Never present a flat 0.1/0.2/1.0 triad without the dose bands it belongs to.
Domein 3: Residuen – Tegenionen, Oplosmiddelen, Nitriet en de nitrosaminekwestie
Residual testing for a synthetic peptide has to cover three synthesis-specific residue classes before it can answer the nitrosamine question: the counterion left by purification, the solvents carried through synthesis and cleavage, and residual nitrite introduced with excipients or water. A nitrite result is a screening input, not a release result, and treating the two as the same thing is the most common error in this domain.
Counterion and solvent control is the better-characterized half. Trifluoroacetate from RP-HPLC purification is the standard counterion concern, and volatile organic impurities including DMF, acetonitril, TFA, DCM and NMP are quantified by GC-MS against the ICH Q3C class limits. Nitrite is harder, because it arrives with the formulation rather than the API. Vendor-compiled excipient nitrite testing data place common excipients in the single-digit ppm range or lower, though those figures come from industry testing programs rather than from any agency limit.
Opmerking:Excipient nitrite ppm values cited here are vendor- and industry-derived testing data, not regulatory limits. Confirm them against your own supplier’s data before using them in a specification.
Implementation has three parts. Screen incoming excipients and purified water for nitrite at the method’s limit of quantification. Run a nitrosamine risk assessment for peptides that traces each nitrosamine-forming source category FDA’s nitrosamine control guidance recognizes, including secondary and tertiary amines, quaternary ammonium salts, and nitrite salts. Then define the decision point in advance: a nitrite result above the screening threshold escalates to confirmatory nitrosamine testing rather than being reported as one.
The failure mode is a risk assessment closed without a documented source review, or a nitrite figure filed as a nitrosamine result. Both leave the file unable to show why the conclusion was reached.
Domein 4: Afbraakproducten en stabiliteitsindicatief methodeontwerp
A stability-indicating method is only as good as the degradation pathways it was designed to see. Voor peptiden, that means oxidation, deamidatie, aggregation and truncation, each with its own mechanism and its own chromatographic signature. Design against a generic impurity panel and the method will report a clean profile whether or not the molecule is degrading.
Peptide-pegyleringThe implementation order matters. Run forced degradation pathway by pathway, under conditions that stress one mechanism at a time, then confirm each degradant peak by mass and peak purity rather than assuming co-elution is absence. The industry analysis of the nitrosamine guidance shift notes that peptide nitrosamine strategy now belongs inside forced degradation and stability-indicating purity methods, not in a separate nitrosamine study bolted on later. Which degradants then need formal identification and qualification is a phase-appropriate decision: tighter at later phases, and documented as a decision rather than left implicit.
De faalmodus is stil. A stability program that shows no degradation may simply be running a method that cannot resolve a co-eluting truncation product, so a passing purity result reflects method blindness rather than a stable molecule. That is the gap that blocks formulation lock, because the data cannot support the claim.
Peptide drug product characterization before formulation therefore starts with pathway coverage, not with a number.
Domein 5: Interfaces voor verpakkingen en containersluiting
Container closure and packaging interface testing for peptides belongs in the readiness framework because the container is a contact material, not a procurement line item. FDA’s nitrosamine control guidance names nitrocellulose-containing blister foil as a recognized source of nitrite that can migrate into a drug product, which means a packaging decision can change the impurity profile of the finished peptide. A packaging-interaction analysis published in 2026 describes the same mechanism for elastomer accelerators and PVC/PVdC secondary packaging, and while that analysis is industry commentary rather than a regulator position, the direction of travel is consistent with the FDA guidance.
Implementation starts with scoping leachables screening to the materials that actually touch the drug substance or drug product, then reviewing extractables data before the packaging configuration is locked. USP <1663> extractables framework sets out how to design that assessment, and the companion leachables chapter, USP <1664>, covers the safety-qualification step. ISPE’s vial container-closure integrity review adds the physical-integrity dimension for vials and stoppers.
The failure mode is a packaging change after formulation lock: a new stopper, a different foil laminate, or a switch from glass to a polymer container can introduce a leachable or nitrite source that invalidates the impurity profile the release specification was built on. Depsipeptiden
Contact material
Interface risk
Test that addresses it
Nitrocellulose-containing blister foil
Nitrite migration into drug product
Extractables screening with nitrite-specific detection
Domein 6: Documentatie en gegevensintegriteit — Het bewijsmateriaal controleerbaar maken
A peptide analytical readiness framework is only as strong as the records behind it. Documentation is not a submission-stage assembly task: the same data-integrity expectations that govern manufacturing records apply to the analytical data itself, which is why ALCOA (toerekenbaar, leesbaar, gelijktijdig, origineel, nauwkeurig) is the right lens for a release package.
Three elements make a package auditable. Eerst, a certificate of analysis whose scope matches the methods actually run, so purity, identiteit, and residual results are traceable to a named procedure rather than a generic claim. Seconde, method validation packages aligned to the ICH Q2(R2) validatie raamwerk, which sets out the current expectations for analytical-procedure validation, including how each parameter was demonstrated for the intended use. Derde, nitrosamine and NDSRI reporting that states the tested limit and the method’s sensitivity, not just a pass result.
A pre-formulation checklist that maps each of the six domains to its evidence is the practical way to hold this together. Some synthesis and analytical services providers, MOL Changes among them, supply method qualification packages and impurity libraries that can be used to populate parts of that checklist; the same evidence can be assembled from other partners or generated in-house.
The failure mode is a data package that answers the question asked but cannot survive a documentation review: results without method context, or a CoA scope narrower than the release decision it is meant to support.
Aan de slag: De eerste drie stappen vóór de formuleringsvergrendeling
Start with a gap inventory, not a new test. Take the six domains in this peptide analytical readiness framework and mark, for each one, what evidence already exists and what is missing. Most teams find that identity and purity data exist in some form while residuals, degradation and container-closure evidence do not, which tells you where the first real work sits.
Step two is the nitrite and nitrosamine source review. Walk the excipients, the water system and the packaging components, and document the outcome even when it comes back negative. A negative result with a written method and a date is evidence; an unwritten one is an assumption that has to be rebuilt later under time pressure.
Step three is the reference-standard and method-qualification plan. Decide now which identity and purity methods will carry the program, so that evidence is generated once, at the right phase, against a qualified method rather than re-run against a moving target after formulation lock.
The common hesitation is that this is work for later. The cost of a gap found after formulation lock is not the test itself; it is the reformulation, the repeat stability work and the documentation rebuild that follow a change to the drug product.
Veelgestelde vragen
Betekent een nitrietsignaal dat nitrosamine aanwezig is??
Nee. A nitrite result is an input to a nitrosamine risk assessment for peptides, not a finding of nitrosamine itself. Nitrite still has to meet a secondary amine under conditions that favour nitrosation, and any resulting NDSRI has to survive purification and storage at a detectable level. Treat a positive result as a trigger to run the formation and carryover assessment, not as a release failure.
Welke aanvaardbare innamelimiet geldt, en waarom verschillen de autoriteiten van mening??
The limit follows the authority releasing the material. FDA’s NDSRI acceptable-intake guidance sets a default acceptable intake for nitrosamine drug substance-related impurities, while Health Canada’s lot-by-lot release expectation applies its own limit and release conditions. The difference is scope and legal basis rather than disagreement about the chemistry, so the applicable limit follows the market you release into.
Zijn de ICH Q3A- en Q3B-drempels van toepassing op peptiden??
Gedeeltelijk. ICH Q3B(R2) degradation-product thresholds were written for small-molecule drug products, and EMA’s synthetic-peptide guideline addresses how synthetic peptides should be controlled instead. Teams commonly use the Q3B reporting, identification and qualification thresholds as a familiar reference point for peptide impurity profiling and release testing, then justify peptide-specific limits against the EMA guideline and the actual impurity profile.
Wat doet een 0.1% kwantificeringslimiet gemiddelde in de praktijk?
It means the method can measure a related substance at 0.1% of the main peak, so any impurity at or above that level is quantified rather than merely observed. What sits underneath the number decides whether it holds: resolution from neighbouring peaks, a demonstrated signal-to-noise ratio at that level, and a validated range that covers it. Een methode die rapporteert 0.1% without resolving a co-eluting truncation reports a passing purity that is not real.
Wanneer is het testen van de containersluiting en de verpakkingsinterface vereist??
When the container or delivery system can change what the patient receives. Container closure and packaging interface testing for peptides is required where the formulation contacts a surface that may leach, adsorb, or interact with the drug product, en USP <1663> extractables framework is the standard starting point for designing that assessment. Early development can often justify a documented risk evaluation in place of full testing; later phases and any change of container or supplier generally cannot.
Wat hoort er thuis in een methodevalidatiepakket??
A method validation package should let a reviewer reproduce your conclusion without asking you a question. Onder de ICH Q2(R2) validatie raamwerk, that means the protocol and report, specificity and forced-degradation data, accuracy and precision across the working range, the quantification and detection limits with the basis for each, solution stability, and the system suitability criteria the routine method will be judged against. The gap most often found in review is missing traceability: results that cannot be tied back to the protocol version that produced them.
Hoe veranderen fase-geschikte drempels tussen vroege ontwikkeling en klinische levering??
They tighten as the phase advances and patient exposure grows. Early development can set thresholds around what the method can measure and what the process can control, with the reasoning documented. By clinical supply, thresholds need to be justified against the EMA guideline, supported by stability-indicating data, and consistent with the release specification the receiving authority will see. The framework does not change between phases; the evidence required to defend each threshold does.
Conclusie
A nitrite signal is an entry point, geen oordeel, and the peptide analytical readiness framework treats it that way: six domains evaluated together, because a passing result in one rarely survives a gap in another. Identiteit, profilering van onzuiverheden, residuen, degradation behavior, packaging interfaces and documentation all feed the same release decision, and the weakest domain sets the ceiling on what the data can support.
The regulatory baselines cited here are current as of writing and will move. Nitrosamine limits, solvent classifications and compendial methods are revised on their own schedules, so treat any threshold in this article as a starting reference to confirm against the current primary document before you lock a specification.
Where to start is the gap inventory, not the next assay. Map what you can already demonstrate across all six domains, mark what you cannot, and let that map order the work before formulation lock.
If a difficult sequence or an incomplete documentation set is the constraint, you can request the analytical documentation package or discuss the sequence directly with the team.
Openbaring: this article is published by a commercial peptide synthesis and analytical services provider, and the framework described reflects capabilities offered commercially.
Hoofd Technologie; Expert op het gebied van peptidesyntheseKernexpertise: Complexe peptidesynthese, niet-natuurlijke aminozuurmodificaties, en de constructie van cyclische peptiden en geniete peptiden.
Biografie:Zejun Peng heeft uitgebreide ervaring in organische chemie en peptidesynthese. Hij is bedreven in de gecombineerde toepassing van vaste-fase-peptidesynthese (SPSS) en peptidesynthese in de vloeibare fase (LPPS), en is bijzonder bedreven in het overwinnen van “extreem moeilijk te synthetiseren sequenties” (zoals peptiden met ultralange keten, zeer hydrofobe sequenties, en meervoudige vouwing van disulfidebindingen). Onder zijn leiding, het team heeft met succes technische knelpunten overwonnen in verschillende gespecialiseerde aanpassingen (zoals N-methylering, PEGylatie, en fluorescerende etikettering), het handhaven van een succespercentage van de synthese van meer dan 98%.
Hallo daar! 👋 Welkom bij MOL Veranderingen. Hoe kunnen wij u vandaag helpen? Vraag gerust naar onze peptidesynthese, CRO-diensten, or any product inquiries — just type your message below and we'll continue the conversation on WhatsApp.