What is the intended use and risk classification of an AI tool in peptide QC?
M Peptides Peptide Hgh Classification follows what the output does to the record, not what the vendor calls the tool. The FDA’s January 2025 draft guidance asks sponsors to define a context of use covering the regulatory question, the model’s role and scope, intended users, inputs and outputs, how outputs combine with other evidence, and whether the model decides, recommends for human review, or is one input to a decision (FDA draft guidance). The same draft sets a risk-based credibility framework: sponsors weigh the model’s influence on the decision and the consequences of an incorrect output, then scale data quality, validation and testing, performance metrics, robustezza, bias, error mitigation and explainability accordingly (FDA press announcement, 2025-01-06).
Two status facts change how much weight that framework carries. The draft had not been finalised as of CDER’s page update, which still lists it as a 2025 draft guidance (CDER, content current 2026-05-01). The ten principles FDA and EMA released on 14 Gennaio 2026 are voluntary, are not a binding rule, and are not a replacement for the draft (FDA/EMA Guiding Principles).
Peptides add a scope problem. ICH Q6B excludes synthetic peptides and polypeptides from its scope, so a peptide lab cannot inherit an upstream purity specification from that guideline (Io Q6B, Fare un passo 4, 1999-03-10). The obligations that do bind are older and already in force: Parte 11 requires system validation “to ensure accuracy, affidabilità, prestazione prevista coerente, and the ability to discern invalid or altered records” (21 CFR 11.10(UN)), and Annex 11 states that “the application should be validated; IT infrastructure should be qualified” (Allegato 11 Principio).
What makes the input data suitable and traceable for an AI model in a GxP lab?
Data suitability is a documented property, not an assumption. The MHRA defines data integrity as “the degree to which data are complete, coerente, accurato, trustworthy, reliable and that these characteristics of the data are maintained throughout the data life cycle,” and adds that “there is no difference in expectations regardless of which acronym is used” between ALCOA and ALCOA+ (MHRA GXP Data Integrity Guidance). The same guidance states that metadata “form an integral part of the original record” and that raw data must permit full reconstruction of the activity. Applied to a model, the inputs, the model version and its parameters are part of the record, not optional context.
The adoption picture is a forecast, not an outcome. In a 2025 Deloitte survey of 103 biopharma QC executives, 50% reported fewer errors and deviations, 45% improved compliance and 43% shorter testing timelines, while 38% expect to embed AI and machine learning in QC operations within two to three years (Deloitte 2025 QC Lab of the Future Survey, fieldwork April 2025). Those are self-reported executive figures with geography not stated, not measured cycle times, and the survey’s own note says the sample may not represent the whole industry.
The counter-condition is the one that matters for traceability: 40% of the same executives said their QC labs still operate with disconnected systems and limited automation, with data fragmented across LIMS, electronic lab notebooks and individual instruments (Deloitte survey). Fragmented systems are exactly the condition under which audit trails and model traceability are hardest to demonstrate.
Industry GAMP guidance can inform input logging, prediction traceability and drift monitoring. Treat it as industry good practice, not an independent legal obligation; connect each control to intended use, applicable requirements and a documented risk assessment.
What exactly gets validated, and against what predefined acceptance criteria?
Two things get validated, and generic guidance merges them. The analytical procedure is validated under ICH Q2(R2), which requires that “prior to the validation study, a validation protocol should be generated,” containing the intended purpose of the procedure and the performance characteristics and associated criteria to be validated (Io Q2(R2)). Precisione Aziende di peptidi and precision may be evaluated independently, each against a predefined acceptance criterion. The model is validated separately, against the context of use and the credibility activities the FDA framework scales to the model’s influence on the decision (FDA draft guidance).
Impurity acceptance criteria require a product- and pathway-specific rationale. ICH Q6A and Q6B do not supply a universal synthetic-peptide specification; their principles may inform an assessment only within the appropriate scope. Use applicable peptide guidance, qualified impurity levels, process capability and stability evidence.
What a documented model metric looks like is available, but only as an example. A deep-learning chromatographic integration model was developed to a target of greater than 90% accuracy and fewer than 10 A 15% integration errors, with retention-time prediction within 0.3 minutes for 95% of IEX peaks and boundary errors of roughly 0.1 A 1.3 minutes depending on method (Biotechnology and Bioengineering, 2024). No source connects figures of that kind to a regulatory acceptance threshold, so they illustrate the metric a sponsor would document, not a benchmark to copy.
The failure mode is a protocol written after the data. Allegato 11 requires manufacturers to justify “standards, protocolli, criteri di accettazione, procedures and records based on risk assessment,” and to include change-control Wholesale Peptide Supplier records and reports of deviations in the validation documentation (Allegato 11 §4.1, §4.2).
How are AI models versioned and change-controlled over time?
Versioning is a controlled-system requirement, not a software convenience. Allegato 11 §10 states that “any changes to a computerised system including system configurations should only be made in a controlled manner in accordance with a defined procedure,” and §11 requires periodic evaluation to confirm the system remains in a valid state (Allegato 11). A retrained model is a changed system under that wording.
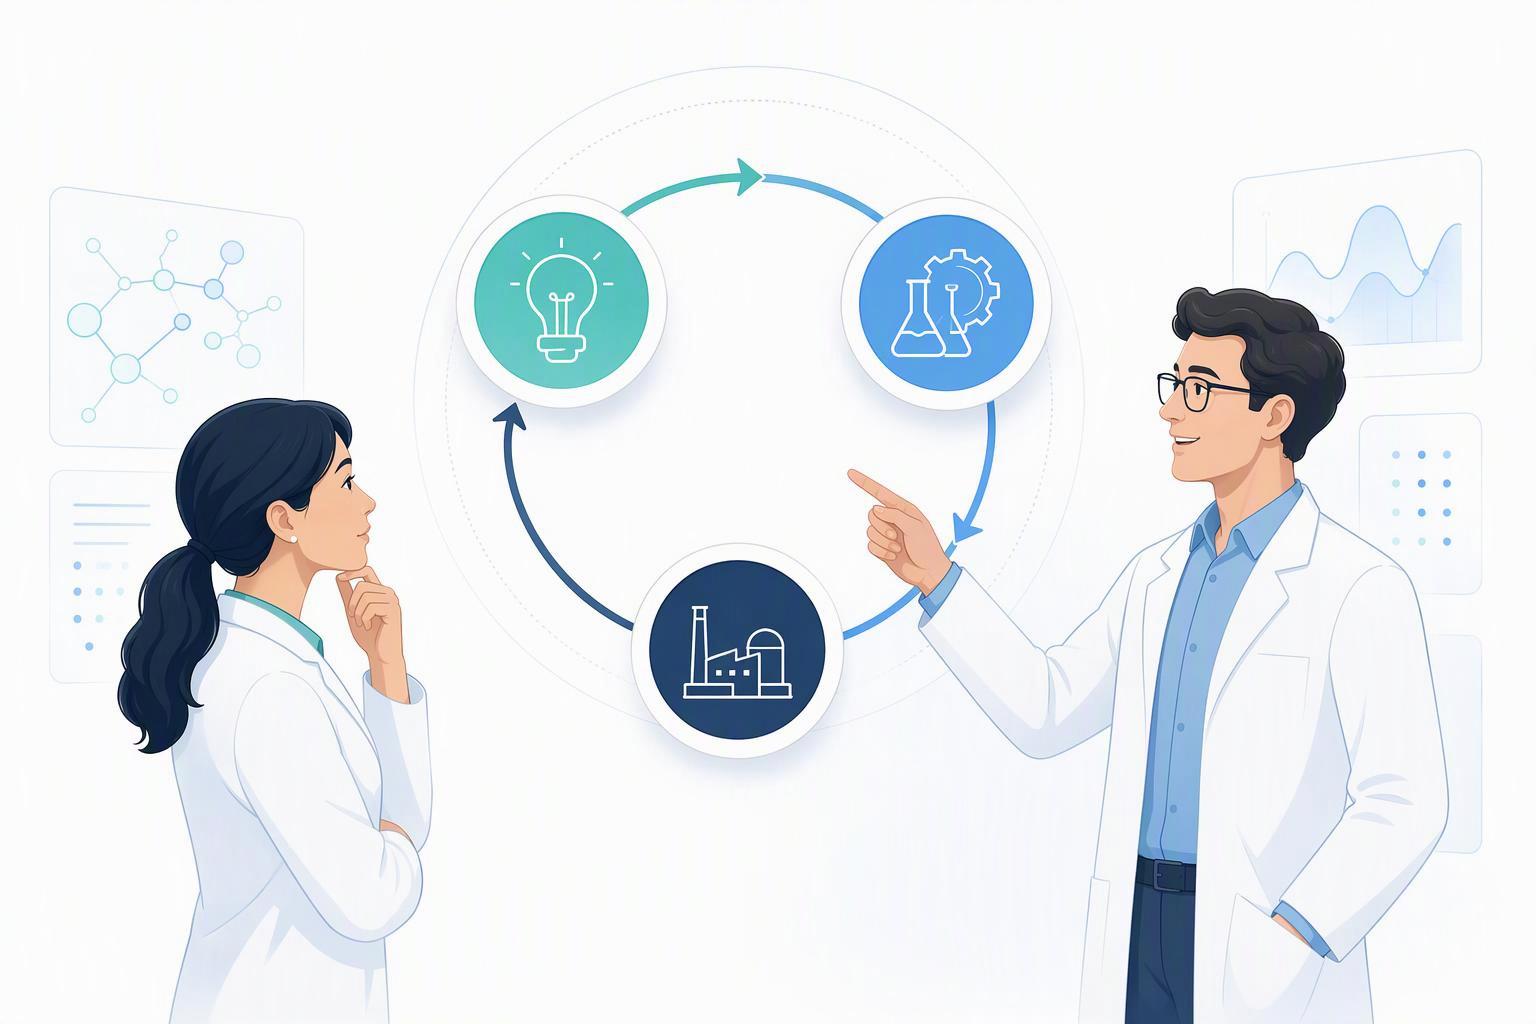
GAMP 5 Second Edition describes the machine-learning subsystem lifecycle in three phases: Concept (destinazione d'uso, GxP context, risks, performance objectives, criteri di accettazione), Project (iterative data preparation, model selection, training, tuning, testing against predefined criteria, with training, validation and test data separated), and Operations (controlled deployment, performance monitoring, data and model governance, incident handling, periodic review, controlled updates) (ISPE GAMP 5 Second Edition, Luglio 2022). The lifecycle is iterative: operational monitoring can send a model back into assessment, development or revalidation.
ISPE’s GAMP guide on artificial intelligence makes the triggers concrete. Retraining is governed by predefined criteria, either performance below an established threshold or the arrival of new data, and continuous training and retesting should be considered when data are added, algorithms fine-tuned or a model retrained, with a documented impact assessment before a changed model is released (ISPE GAMP Guide: Artificial Intelligence, 2025). Model versioning exists so changes are traceable and results reproducible.
The failure mode is a model update that was never re-qualified. Parte 11 requires revision and change-control procedures that maintain an audit trail documenting the time-sequenced development and modification of systems documentation (21 CFR 11.10(k)), and Annex 11 expects change-control records inside the validation documentation itself.
Where does human review remain mandatory when AI touches a QC decision?
Assign accountable quality-unit or Qualified Person responsibilities under the applicable GMP framework. Parte 11 authority and signature controls do not themselves require manual approval of every intermediate AI output. Validate the automated steps, escalation criteria and information used in the final disposition.
The EU AI Act supplies the oversight vocabulary where a system qualifies as high-risk: such systems must be designed so they “can be effectively overseen by natural persons during the period in which they are in use,” and oversight must let the overseer understand capabilities and limits, remain aware of automation bias, interpret outputs correctly, decide not to use or to override or reverse the output, and intervene or stop the system (EU AI Act Article 14). Article 14 applies to Annex III high-risk systems from 2026-08-02 and to Annex I embedded high-risk systems from 2027-08-02. Pharma QC AI is not automatically high-risk; the classification depends on the product function, so the condition has to be assessed rather than assumed.
The FDA framework reaches the same place from the model side: the context of use must state whether the model decides, recommends for human review, or is one input to a decision (FDA draft guidance). No source establishes a universal confidence threshold, such as 0.80 O 0.95, for routing an output to human review. That absence is the point the risk-based framework makes: the threshold must be justified and validated for the intended use.
La modalità di guasto è documentata. UN 2025 FDA warning letter cited a firm under 21 CFR 211.68(B) because, during review of analytical system audit trails for final release testing, the inspection found the firm did not review audit trails and raw analytical data captured by its instruments and had no written procedures to review electronic data or integration for batch release (Wisconsin Pharmacal Company, warning letter 320-25-103, 2025-08-22). UN 2021 letter to Adamson Analytical Laboratories states that “21 CFR 211.22 requires data review prior to batch release including audit trail data” (Adamson Analytical Laboratories, warning letter 614644, 2021-08-17).
What does an auditor expect to see in an audit-ready AI validation record?
An audit-ready record is a set of named fields, not a narrative. Parte 11 richiede sicurezza, generato dal computer, time-stamped audit trails that independently record the date and time of operator entries and actions creating, modificare o cancellare record elettronici, states that record changes “shall not obscure previously recorded information,” and requires retention at least as long as the subject records and availability for agency review and copying (21 CFR 11.10(e)). Signatures must carry the printed name of the signer, the date and time, and the meaning of the signature, such as review, approval, responsibility or authorship (21 CFR 11.50).
Allegato 11 words the audit trail requirement differently: consideration should be given, based on a risk assessment, to building in a record of all GMP-relevant changes and deletions, the reason for a change or deletion should be documented, and audit trails need to be available, convertible to a generally intelligible form, and regularly reviewed (Allegato 11 §9). Parte 11 is prescriptive; Allegato 11 is risk-gated. A lab designing one control set for both markets has to satisfy the stricter reading.
The MHRA vocabulary tells the auditor what completeness means for a model record: data must be attributable, leggibile, contemporaneo, originale e accurato, and governance measures must keep it complete, coerente, enduring and available throughout the lifecycle, with metadata forming an integral part of the original record (MHRA GXP Data Integrity Guidance). For an AI tool, that means the model version, its inputs, its prediction and the reviewer’s action all sit in the same reconstructable record.
The FDA framework’s credibility activities define what the record has to demonstrate about the model itself: data quality, validation and testing, performance metrics, robustezza, bias, error mitigation, and explainability where needed, scaled to the model’s influence and the consequences of an incorrect output (FDA press announcement, 2025-01-06). The ten principles FDA and EMA published in January 2026 add data governance and documentation, risk-based performance assessment, life cycle management, and clear, essential information as named expectations (FDA/EMA Guiding Principles).
One gap should be stated plainly: no source describes how a specific AI model satisfies Part 11’s signature-to-record linking, the uniqueness of electronic signatures, or the two-distinct-identification-component requirement, and none describes how a model interacts with Annex 11 §15, which allows only Qualified Persons to certify batch release using an electronic signature (Allegato 11). The sources do not address it, so the validation file has to. Wholesale Peptide Factory
How does AI validation evidence connect to the CoA and lot-to-lot package a peptide buyer reviews?
The CoA is the downstream artifact of the validation record, and its purity line is only as strong as the method behind it. Q6B states that purity is “highly method-dependent” and that “the purity of the drug substance and drug product is assessed by a combination of analytical procedures” (Io Q6B). A single purity percentage on a peptide CoA therefore carries the method identity, the acceptance criterion and the validation status of that method with it, or it carries nothing an auditor can check. Peptide Technologies
Compare lot results against justified criteria for the actual product, method and impurity risk. A simple mean-plus-three-standard-deviations calculation neither establishes safety qualification nor supplies a universal synthetic-peptide release limit. Document the scientific and regulatory basis for each criterion.
Where an AI tool touches the release path, the CoA package inherits the electronic-record obligations. Parte 11 requires the ability to generate accurate and complete copies of records in human readable and electronic form suitable for inspection, review and copying, and requires protection of records for accurate and ready retrieval throughout the retention period (21 CFR 11.10(B), (C)). Allegato 11 §8.2 adds that for records supporting batch release it should be possible to generate printouts indicating whether any data has been changed since the original entry (Allegato 11).
The scale of the review problem is visible in published triage data. In a 2024 Microchemical Journal study, a supervised-learning peak evaluation and automated review tool handled 85% of chromatogram segments automatically and sent the remaining 15% for manual review (Alewijn et al., Microchemical Journal 207:111578, 2024). That is a triage rate, not a false-positive rate, and the study reported equivalent quantitative performance to expert integration. A vendor-reported figure for a different tool claims roughly 90% reproduction of expert integrations and 88% correct peak detection against 48% for the conventional algorithm (Shimadzu Peakintelligence for LC, vendor-reported, 2024-11-01); the two figures measure different things and should not be compared directly.
Conclusione
The evidence pack for AI validation in peptide quality operations has six boxes, and none of them requires a peptide-specific precedent to fill. Define the context of use and classify the tool by what its output does to the record, per the FDA’s January 2025 draft guidance (FDA draft guidance). Document data suitability in MHRA terms, with metadata as part of the original record (MHRA GXP Data Integrity Guidance). Write the validation protocol and its predefined acceptance criteria before the study, under ICH Q2(R2) (Io Q2(R2)). Version and change-control the model as a controlled system under Annex 11 §10 and the GAMP lifecycle (Allegato 11; ISPE GAMP 5 Second Edition). Keep human review and its override authority in the record under Part 11 §11.10(G) and §11.70 (21 CFR parte 11). Produce audit trails that do not obscure prior data and are regularly reviewed (Allegato 11 §9).
Two honest gaps remain. No source documents an AI tool making or supporting a polypeptide release decision, and no regulator has specified a confidence threshold for routing AI output to human review. Both gaps are the reason the validation file, not the vendor’s claims, is the evidence an inspector will read.
Start with one defined use case. Document its regulatory question, data inputs, model version, criteri di accettazione, review responsibilities and change triggers, then assess the evidence gaps before deployment.