Khung quản trị nghiên cứu peptide được hỗ trợ bởi AI là gì?
Khung quản trị nghiên cứu peptide được hỗ trợ bởi AI là bộ quy tắc, hồ sơ, và xem xét các cổng giúp thiết kế do AI soạn thảo có thể theo dõi được một người có tên có thể bảo vệ nó. Nó nằm cạnh các hệ thống chất lượng mà phòng thí nghiệm đã vận hành, không ở trên họ, và nó tồn tại vì bốn hệ thống phân bổ khác nhau hiện chồng chéo lên nhau trên cùng một công việc.
Bốn hệ thống đó trả lời các câu hỏi khác nhau và được quản lý bởi các cơ quan có thẩm quyền khác nhau. Hướng dẫn về quyền phát minh sửa đổi của USPTO nêu rõ rằng chỉ thể nhân mới có thể được nêu tên chính xác là nhà phát minh, và coi hệ thống AI là công cụ được các nhà phát minh con người sử dụng theo cùng tiêu chuẩn pháp lý áp dụng cho mọi phát minh khác. Phân bổ theo quy định chạy trên một kênh riêng biệt: Phần của FDA 11 hướng dẫn về phạm vi và ứng dụng áp dụng cho hồ sơ điện tử được tạo, đã sửa đổi, duy trì, đã lưu trữ, đã lấy lại, hoặc được truyền đi theo yêu cầu về hồ sơ theo quy định của FDA, với những yêu cầu cơ bản đó đóng vai trò là quy tắc vị ngữ.
|
Hệ thống phân bổ |
Điều gì nó quyết định |
Ai có thể giữ nó |
AI có thể giữ nó được không? |
Ghi lại bằng chứng cần thiết |
|---|---|---|---|---|
|
quyền giám hộ |
Ai giữ tài liệu và khi nào |
Người hoặc trang web được đặt tên |
KHÔNG |
Nhật ký chuỗi hành trình sản phẩm, mục lưu trữ và chuyển giao |
|
Quyền tác giả |
Ai viết lý lẽ và phân tích |
Người được nêu tên |
KHÔNG |
Mục nhập ngày tháng, hồ sơ nhắc nhở và đầu ra, đăng xuất |
|
phát minh |
Ai đã nghĩ ra phát minh được yêu cầu bồi thường |
Chỉ thể nhân |
Tổng hợp peptit KHÔNG |
Hồ sơ thụ thai, tường thuật đóng góp |
|
Phân bổ theo quy định |
Ai chịu trách nhiệm trước cơ quan |
Pháp nhân và cá nhân chịu trách nhiệm |
KHÔNG |
Bản ghi quy tắc vị ngữ, dấu vết kiểm toán, chữ ký điện tử |
Từ ngữ ở đây tuy vô duyên nhưng lại có sức nặng. ALCOA+ mô tả chất lượng bản ghi: có thể quy cho, dễ đọc, đương thời, nguyên bản, chính xác, cộng với hoàn thành, nhất quán, bền bỉ, và có sẵn. ELN là sổ tay thí nghiệm điện tử; LIMS là một hệ thống quản lý thông tin phòng thí nghiệm; một phương pháp trực giao xác nhận kết quả bằng nguyên tắc độc lập về mặt hóa học thay vì lặp lại cùng một phép đo.
Peptide tổng hợp c=”https://molchanges.com/wp-content/uploads/2026/09/pub_20260921_225551_664_f7182fac5c724f199d0c23c32516026d.png”>
Quyền sở hữu bản ghi thử nghiệm và tính toàn vẹn dữ liệu nằm ở điểm giao nhau của cả bốn hệ thống, đó là lý do tại sao khung phân tách chúng trước khi yêu cầu bất kỳ ai thay đổi quy trình làm việc. Quan niệm sai lầm phổ biến là quản trị làm chậm tiến độ khoa học. Trong thực tế, nó loại bỏ công việc tái thiết sau một kết quả gây tranh cãi, và việc tái thiết đó là phần chậm.
Tại sao quản trị nghiên cứu peptide được hỗ trợ bởi AI lại quan trọng hiện nay
AI sáng tạo đã chuyển từ tính mới sang sản xuất trong hóa học, và hồ sơ bằng sáng chế cho thấy điều đó. Tháng 7 của WIPO 2026 Số lượng cập nhật bằng sáng chế GenAI xấp xỉ 14,000 các họ bằng sáng chế AI mang tính sáng tạo trong 2023 và hơn thế nữa 37,800 TRONG 2025. Phân tích bằng sáng chế GenAI của WIPO đặt 2024 hình dung tại 18,862 và 2025 hình dung tại 37,808, so filings roughly doubled in a single year.
Regulators moved at the same time. các 2025 revision of the USPTO’s inventorship guidance rescinds the February 2024 AI-inventorship guidance in full and implements Executive Order 14179.
The consequence of ignoring this is concrete. An experimental record assembled from a chat thread and an unlabelled aliquot cannot support an inventorship claim, a reproducibility claim or a safety claim. This is where AI-assisted peptide design documentation stops being paperwork and becomes the asset itself.
thuốc 1: Ghi lại lời nhắc và lý do thiết kế
Documenting prompts and design rationale means capturing the AI interaction and the human judgement that followed it as one linked record, not two separate artefacts. The prompt alone proves nothing: it shows what the model returned, not why a scientist advanced one candidate and rejected the rest.
The field list below is common practice drawn from software-vendor prompt-management tooling. Only the record attributes are grounded in the regulatory cluster. A workable log entry carries the model name and version, the prompt text as sent, parameters and seed, a timestamp, người điều hành, the candidate set returned, and a written rationale for each accept or reject decision. Each field then maps to the electronic laboratory notebook (ELN) entry it must accompany, so the design record and the experimental record stay joined.
That mapping matters because of how regulators read the rules. FDA diễn giải Phần 11 scope narrowly: it applies to records required under predicate rules that are kept electronically in place of paper, or kept electronically in addition to paper and relied on to perform regulated activities (FDA, Phần 11 scope and application, page current as of 2018-08-24). các MHRA’s GxP data-integrity guidance supplies the attributes that make such a record defensible: ALCOA+ (Có thể quy cho, Dễ đọc, Đương thời, Nguyên bản, Accurate, plus Complete, Nhất quán, Bền bỉ, Có sẵn), and an audit trail defined as metadata that allows reconstruction of a record’s history, including who, what, when and why.
Chế độ thất bại quen thuộc. A design rationale that lives only in a chat thread cannot be reconstructed when inventorship is challenged, because the human contribution is invisible. How much of the prompt chain you retain scales with program stage: research-grade work can keep the final prompt and rationale, while GMP-bound material needs the full chain, including rejected candidates.
thuốc 2: Cổng đánh giá con người cho nghiên cứu được hỗ trợ bởi AI
A review gate is a decision point with a named owner, a defined rejection criterion and a route for appeal. It is not a meeting where a draft is discussed and then quietly advanced.
The distinction matters because the accountability question is already settled in law. The USPTO’s position is that AI systems are tools used by human inventors, và USPTO’s AI-inventorship FAQs confirm the analysis stays focused on human contribution: the guidance “does not create a heightened standard for inventorship” and adds no separate AI-disclosure duty beyond existing rules (USPTO, 2025-01-14). Whatever the model proposed, a person owns the decision.
Dịch vụ Four gates carry that ownership in practice.
|
Cổng |
Reviewer competence |
Sign-off authority |
Rejection criteria |
Escalation path |
|---|---|---|---|---|
|
Candidate advancement |
Domain scientist familiar with the target and prior art |
Principal investigator or project lead |
Rationale not reproducible from the prompt log; no orthogonal rationale for the proposed modification |
Program director, with the prompt log attached |
|
Wet-lab commitment |
Chemist or process owner who will run or commission the work |
Laboratory manager |
Route not feasible at required scale; reagent or equipment constraints unresolved |
Head of R&D |
|
Phát hành phân tích |
Analyst qualified on Cửa hàng the specific method |
QC lead |
Identity or purity data incomplete for the intended use |
Quality manager |
|
Safety-relevant impurity disposition |
Toxicologist or qualified safety reviewer |
Quality manager, or the qualified person where the material is GMP-bound |
Impurity above the limit set for the intended use, with no justified control |
Quality manager and regulatory affairs jointly |
Who may sign changes with the material’s status. A research-grade reagent can be released on a qualified analyst’s review. A GMP-bound material cannot: release requires the named qualified person under the applicable quality system, and no AI output substitutes for that signature.
Chìa khóa mang đi: A review gate without a named sign-off authority and a defined rejection criterion is not a control, it is a meeting.
Chế độ thất bại yên tĩnh. A gate exists on paper, the discussion happens, and an AI-drafted candidate enters synthesis with nobody accountable for the decision. When the record is later questioned, the trail shows a conversation, không phải là sự chấp thuận.
thuốc 3: Quyền sở hữu bản ghi thử nghiệm và tính toàn vẹn dữ liệu
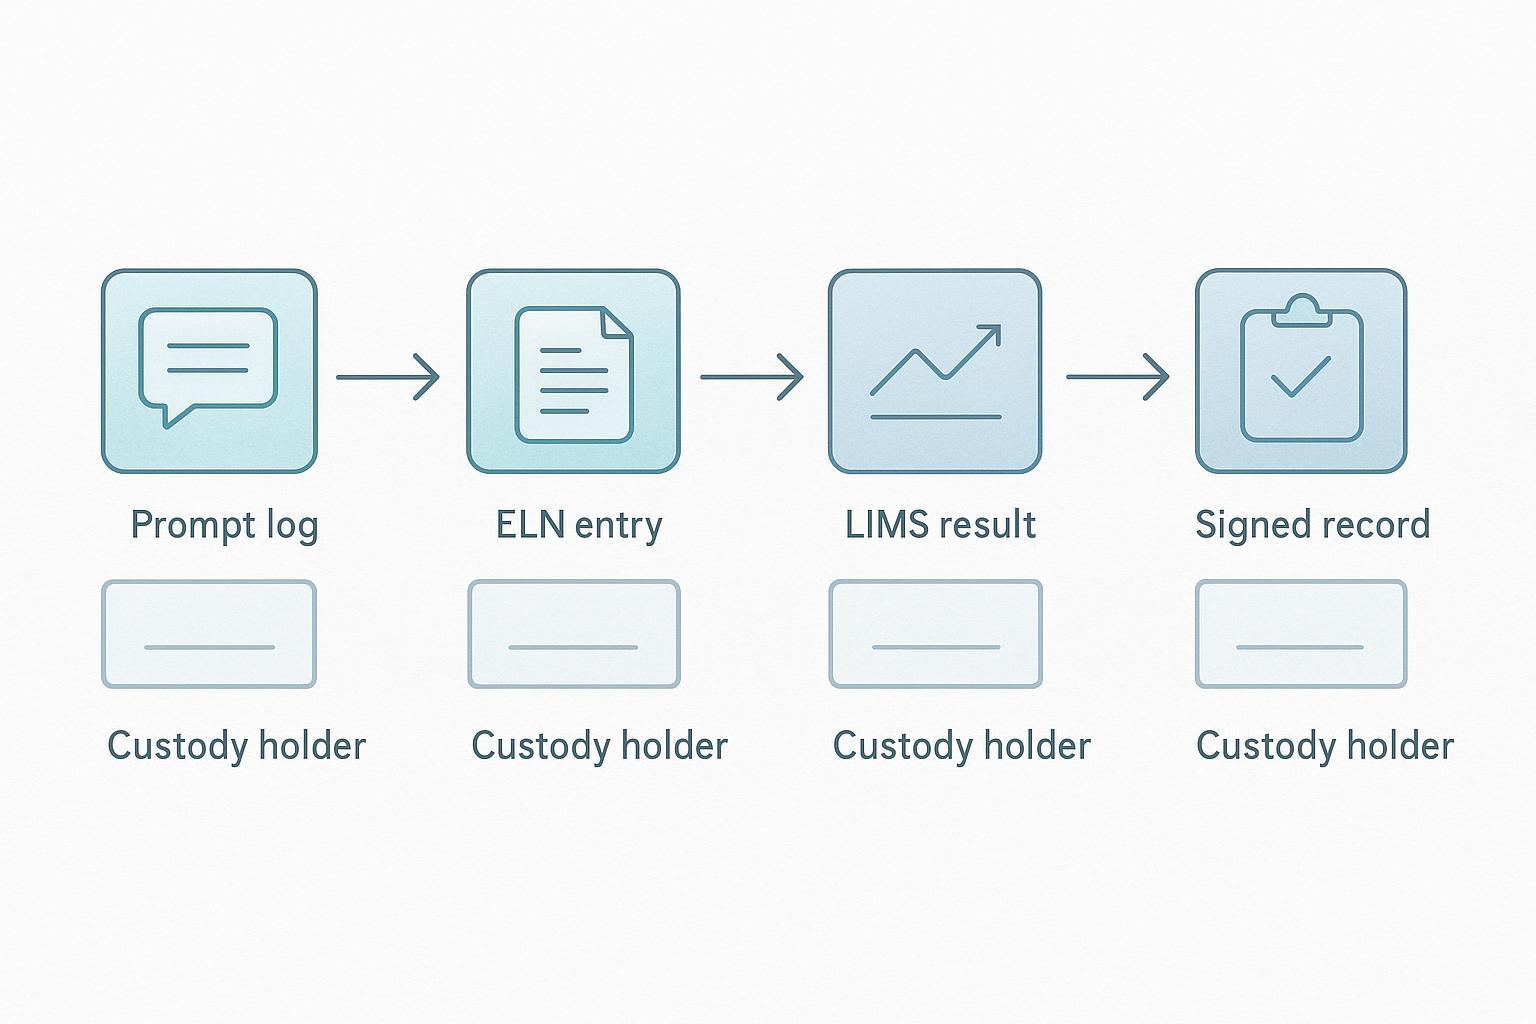
Ownership is four questions, không phải một. Custody asks who physically holds the record; authorship asks who wrote it; inventorship asks who contributed to the invention; regulatory attribution asks who is accountable to an inspector. In most AI-assisted programs those land with four different people, and the record has to satisfy all four at once.
The MHRA’s GxP data-integrity guidance defines ALCOA+ as nine attributes: Có thể quy cho, Dễ đọc, Đương thời, Nguyên bản, Accurate, plus Complete, Nhất quán, Enduring and Available. The same guidance treats the audit trail as metadata capturing who, what, when and why, with every change dated, attributed and justified. FDA’s CGMP definition is narrower in wording and identical in effect: a secure, do máy tính tạo ra, time-stamped record that allows reconstruction of the course of events.
Scope is where teams misread the rules. Phần của FDA 11 scope-and-application guidance attaches the regulation to records created, đã sửa đổi, duy trì, đã lưu trữ, retrieved or transmitted under a records requirement in FDA regulations, and FDA’s narrow reading of Part 11’s scope means paper-equivalent obligations follow the predicate rule, không phải phần mềm.
The failure mode is fragmentation. Dữ liệu thô, intermediate files, analysis scripts and model outputs sit in four places, so no single record reconstructs the run and nobody can say who signed off. Retention and audit-trail review obligations then scale with program stage: a research-grade record tolerates lighter review than a GMP-bound one.
thuốc 4: Xác nhận danh tính peptide
No single analytical method confirms a peptide’s identity. Xác nhận trực giao, meaning two or more methods that rely on different physical or chemical principles, is what turns a plausible result into a defensible identity record for peptide identity purity and impurity confirmation.
Mass spectrometry with MS-MS fragmentation establishes the molecular mass and, through fragment ions, the amino-acid sequence. It cannot tell you whether the material is the correct diastereomer, and it will not distinguish a sequence from a closely related deletion analogue that fragments similarly. Amino-acid analysis establishes the compositional ratio after hydrolysis, which catches substitutions and truncations that mass alone can miss, but it destroys sequence order information and says nothing about chirality. Chromatographic retention against a reference standard establishes that the material behaves identically to a characterised lot under defined conditions, which is a strong similarity statement rather than a structural one.
Run together, the three methods cover each other’s blind spots. Run alone, each leaves a gap that a reviewer or a regulator will find. The method table below sets out what each technique establishes, what it cannot establish, and the program stage at which it becomes mandatory.
|
Orthogonal identity method |
Những gì nó thiết lập |
What it cannot establish |
Mandatory at |
|---|---|---|---|
|
Mass spectrometry with MS-MS fragmentation |
Molecular mass and fragment-derived sequence |
Diastereomer identity; distinction from some closely related analogues |
Research grade onward |
|
Amino-acid analysis |
Compositional ratio after hydrolysis |
Thứ tự tuần tự; chirality |
Research grade onward |
|
Chromatographic retention vs. reference standard |
Behavioural identity to a characterised lot |
Structural identity |
Research grade onward |
The expectations for how these methods are validated sit in Tôi quý 2(R2), the current revision of the analytical-procedure validation guideline, which was adopted at Step 4 and took legal effect in the EU on 14 Tháng sáu 2024, replacing Q2(R1) của 1995. The EMA’s ICH Q2(R2) guideline page records that effective date. Q2(R2) covers validation tests and terminology for procedures used in release and stability testing, and it can be applied to other procedures in the control strategy on a risk-based basis. Instrument suitability is a separate obligation: USP <1058> on analytical instrument qualification requires documented evidence that an instrument is fit for its intended purpose across design, installation, operational and performance qualification.
Chế độ thất bại là cụ thể và phổ biến. A vendor certificate or a model-generated identity summary is filed as the identity record, with no orthogonal confirmation and no instrument-qualification trail behind it. The record then asserts an identity that no one can reconstruct from primary data.
The research-grade versus GMP-bound difference is a matter of degree and evidence. Research-grade work can reasonably rely on a single primary method with a second confirmatory technique where the material’s use justifies it. GMP-bound material raises the bar: the full orthogonal set, each method validated under Q2(R2), and every instrument carrying a current qualification record under USP <1058>.
thuốc 5: Hồ sơ tạp chất liên quan đến độ tinh khiết và an toàn
An HPLC area-percent number is not a purity specification. It reports the share of UV-absorbing signal at 214 hoặc 220 bước sóng, so it cannot see the trifluoroacetate counterion, dung môi dư, elemental impurities, or water carried alongside the peptide. xét nghiệm, counterion content and moisture have to be accounted for together before any purity figure means anything.
The familiar thresholds are convention, not pharmacopeial rule. Suppliers and industry practice commonly quote ≥95% for research-grade material and ≥98%–99% for pharmaceutical-grade, but no compendial chapter fixes those numbers as a pass criterion for a peptide. Treat them as purchasing shorthand and define your own specification against the intended use.
What separates research-grade from GMP-bound material is not the headline percentage but which impurity classes must be quantified rather than merely observed. A research reagent can tolerate a reported counterion and an unspecified solvent profile. A GMP-bound batch cannot: safety-relevant impurities peptide work requires each class to be named, measured against a limit, and dispositioned on the record.
The classes to name and disposition are residual solvents, elemental impurities, trifluoroacetate counterion carryover, deletion and truncation sequences from incomplete coupling, and endotoxin where the route demands it. For parenterals, USP <85>, xét nghiệm nội độc tố vi khuẩn, sets the limit from the maximum human dose per kilogram per hour using the formula K/M, với K = 5 EU/kg/h for most routes and 0.2 EU/kg/h for intrathecal administration (USP <85>). Where sterility is claimed, USP <71> sterility testing requires 14-day incubation in Fluid Thioglycollate Medium and Soybean–Casein Digest Medium, with no growth for a pass (FDA pyrogen and endotoxin testing Q&MỘT).
Automation does not remove this burden. MỘT 2025 review of LLMs in organic synthesis records IBM RoboRXN+ completing a 12-step synthesis of a nonribosomal peptide at 92% sự tinh khiết, and MoleculeX reporting kinase-inhibitor route planning cut from weeks to hours at 70% năng suất (Tharwani et al., 2025). Both figures are area-percent or isolated-yield outcomes. Neither tells you what else is in the vial.
That gap is the failure mode. A 24-mer with a TFA counterion gets carried into a cell assay on the strength of an area-percent number alone, and the assay reads the counterion as much as the peptide. Peptide identity, purity and impurity confirmation are three separate determinations, and only the third one tells you whether the material is fit for the experiment you are about to run.
thuốc 6: Dữ liệu hoạt động và các tạp chất liên quan đến an toàn trong bối cảnh
An activity value means little on its own. A single unreplicated result is a data point, không phải bằng chứng, and the reproducibility literature shows why: TRONG Baker’s 2016 Nature reproducibility survey, nhiều hơn 70% của 1,576 surveyed researchers reported failing at least once to reproduce another scientist’s experiment. That figure is self-reported, not a measured replication rate, but it establishes the practical point. Activity data earn their weight through repetition and through traceability to the material they were generated against.
That traceability is what links activity results to safety-relevant impurities in a peptide program. Every activity value should carry the lot identifier, the purity profile, and the impurity disposition it was measured against, so that a later finding can be traced back to every result it may invalidate. Orthogonal confirmation of peptide identity belongs in the same record, because an activity readout from a misidentified lot is not recoverable after the fact.
The research-grade versus GMP-bound distinction changes both the replication requirement and the threshold at which an impurity finding triggers a re-test. Where a research-grade reagent may tolerate a single determination, a GMP-bound material requires the stricter route-dependent limits described in USP <85>, xét nghiệm nội độc tố vi khuẩn, and any impurity finding above threshold forces affected results back into question.
The failure mode is blunt. An activity result that cannot be tied to the lot it came from leaves no way to scope the damage when a safety-relevant impurity surfaces later, and the whole dataset is discarded rather than selectively re-tested.
Trình độ cao: Mở rộng gánh nặng bằng chứng đến giai đoạn chương trình
If your team already runs an AI-assisted workflow, the insight that matters is this: the AI-assisted peptide research governance framework is not applied uniformly. It scales with patient-safety and quality impact, so a research-grade program that adopts the full GMP-bound package will over-document and stall, while a program heading toward a regulatory submission that adopts the research-grade package will under-document and lose the record.
The scaling rule is simple. Match the evidence burden to the stage: research-grade work needs prompt logs, design rationale, and rejected-candidate retention; development work adds qualified instruments and validated methods; GMP-bound work adds full release and stability testing. The trigger that moves a program up a tier is a regulatory commitment, not a calendar date. Tôi quý 2(R2), the current revision of the analytical-procedure validation guideline, explicitly allows its validation tests and terminology to be applied to other procedures on a risk-based basis, which is the same logic that should govern your documentation depth. USP <1058> on analytical instrument qualification applies the same lifecycle thinking to instruments.
Honestly, this framework is heavier than a pure discovery program needs. Do not scale down two things, mặc dù.
Để biết tiền boa: Never scale down human sign-off authority or rejected-candidate retention. Neither can be reconstructed after the fact, and both are the elements regulators and reviewers ask for first.
Giữ lại ứng viên bị loại và kết quả tiêu cực
A rejected candidate is evidence, not housekeeping. The design that lost, the reason it lost, the prompt and model version that produced it, and the reviewer who turned it down together show the human judgment that selected the winner. Strip those out and the record proves an output existed, but not that a person chose it. That gap is exactly where the question of who owns the experimental record becomes hard to answer.
The retention rule scales with program stage. Research-grade work can keep rejected candidates in an archive with a defined retention period, as long as the rejection rationale stays retrievable. A GMP-bound program should hold them in the controlled record itself, because the selection decision is part of the batch history a reviewer will later reconstruct.
The failure mode is common and quiet: only the winning design survives, so the record reads as if the sequence arrived fully formed. The USPTO’s revised inventorship guidance states that only natural persons may be named inventors, and its AI-inventorship FAQs confirm the analysis turns on human contribution rather than a heightened standard. Rejected candidates are often the clearest proof of that contribution.
Công cụ và cấu trúc bản ghi
Three record structures matter before any software decision: the controlled record system, the prompt and model metadata layer, and the analytical data system. An electronic laboratory notebook (ELN) or laboratory information management system (LIMS) is the controlled record system, and it must carry an audit trail that records who changed what and when. That audit trail is metadata, not a narrative field, and the MHRA’s GxP data-integrity guidance frames it through the nine ALCOA+ attributes: có thể quy cho, dễ đọc, đương thời, nguyên bản, chính xác, hoàn thành, nhất quán, bền bỉ, và có sẵn. The prompt and model metadata layer is the addition most teams still lack, and it must capture the model version, the prompt text, and the human who accepted the output. The analytical data system holds raw instrument files, and it must preserve them unaltered alongside the processed result. Experimental record ownership and data integrity depend on all three connecting, and on one thing none of them supplies: a named sign-off authority. FDA’s narrow reading of Part 11’s scope makes clear that a validated system is not the same as an accountable person.
|
Loại |
Required capability Về |
Typical owner |
Usually already present? |
|---|---|---|---|
|
Controlled record system (ELN/LIMS) |
Audit trail, version history, e-signature |
Quality or data-integrity lead |
Đúng, in a regulated stack |
|
Prompt and model metadata layer |
Model version, prompt text, acceptance record |
Research lead or informatics |
Rarely; usually an addition Sản xuất peptit |
|
Analytical data system |
Raw file retention, unaltered originals |
Analytical or QC lead |
Partly; raw-file retention often needs work |
MOL Changes supports analytical verification and lot traceability as a peptide partner. This is a commercial interest, disclosed here rather than presented as a performance claim.
Bắt đầu
Open the most recent AI-assisted design decision in your program and try to reconstruct it from the record alone: the prompt as it was written, the model and version that produced the output, and the human rationale for accepting or rejecting it. If any of those three is missing, you have found your first gap, and you found it in five minutes without a meeting.
Kế tiếp, name the sign-off authority for the gate ahead and write the rejection criterion down before the work reaches them. A gate with no named owner and no written criterion is a rubber stamp, and the MHRA’s GxP data-integrity guidance is explicit that attributable, contemporaneous records are the baseline, not an aspiration (MHRA GxP Data Integrity Guidance and Definitions, 2018). The same logic runs through inventorship: the USPTO’s revised inventorship guidance confirms that only natural persons may be named inventors, which means an AI system cannot hold the accountability your record implies (USPTO Revised Inventorship Guidance, 2024).
Then pick one pillar and adopt it end to end before adding a second. Prompt traceability is usually the cheapest place to start.
First action: Before you leave this page, open one AI-assisted design decision and check three things: is the prompt recoverable, is the model version recorded, and is the human rationale written down?
The common hesitation is that retrofitting documentation into an existing ELN workflow will disrupt the bench. Starting with a single pillar is how you avoid that: one change, applied completely, tells you what the workflow will tolerate before you commit the program to it.
MOL Changes cung cấp dịch vụ phân tích và tổng hợp peptide tùy chỉnh, vì vậy chúng tôi có lợi ích thương mại trong chủ đề này. If it would help to see how the record structure looks in practice, you can review the documentation model or open a technical discussion with our analytical team.
Câu hỏi thường gặp
Khung quản trị nghiên cứu peptide được hỗ trợ bởi AI là gì?
It is the set of record structures, review gates and sign-off authorities that keep a qualified human accountable for every controlled decision in an AI-assisted peptide workflow. It sits alongside the data-integrity and quality systems a laboratory already runs rather than replacing them, and it does not create a new regulatory regime. các definition section above sets out how the framework maps onto ALCOA+ and 21 Phần CFR 11.
Ai sở hữu hồ sơ thử nghiệm khi công cụ AI phác thảo thiết kế?
quyền giám hộ, quyền tác giả, inventorship and regulatory attribution are four separate questions held by four different parties, and none of them is held by the AI system. Custody decides who can alter the record and when; authorship decides who signed the rationale; inventorship decides who is named on a filing; attribution decides who answers to an inspector. The distinctions table in the definition section assigns each one.
Việc sử dụng AI để thiết kế peptide có ảnh hưởng đến việc ai có thể được mệnh danh là nhà phát minh không?
No separate legal standard exists for AI-assisted inventions, and only natural persons may be named as inventors. The assessment turns on the human contribution to the claimed subject matter, so a designer who set the constraints, selected the candidate and verified the result is assessed the same way as one working without a model. No heightened standard and no new AI-disclosure duty is created; the USPTO rows in the definition section carry the specifics.
Nhật ký nhắc nhở phải chứa những gì trước khi thiết kế được đưa vào ELN?
Model name and version, prompt text, parameters, timestamp and the identity of the person who accepted the output. Without the version and the acceptance record, a reviewer cannot reconstruct why a candidate was carried forward, which is the failure mode Pillar 1 địa chỉ.
Is a vendor’s purity certificate sufficient for a GMP-bound material?
KHÔNG. A certificate establishes what the vendor measured on its own sample; it does not establish identity, and it does not profile safety-relevant impurities the vendor’s method was never designed to see. A 24-mer with a TFA counterion carried into a cell assay is the standard illustration: the counterion and any deletion sequences sit outside a purity percentage. thuốc 4 và trụ cột 5 cover orthogonal confirmation and impurity profiling.
Ứng viên bị từ chối phải được giữ lại trong bao lâu?
Retention follows program stage, and two elements are not retrofittable: the prompt and design rationale, and the identity of the person who accepted the output. A research-grade program can keep rejected candidates in a research archive with the rationale attached; a GMP-bound material needs them inside the controlled record. The scaling section sets out where the line falls.
Chúng tôi có thể lưu trữ các ứng viên bị từ chối thay vì giữ họ trong hồ sơ được kiểm soát không?
For a research-grade program, Đúng, provided the rationale and the accepting reviewer travel with the candidate. The failure mode is an archive that holds the structure but not the reason it was rejected, because the next program repeats the same synthesis.
Phần kết luận
The AI-assisted peptide research governance framework does not settle who owns the experimental record. It makes the human contribution visible enough that the question can be answered at all. Every pillar in this guide serves that single end: documenting prompts and design rationale so the reasoning behind a molecule survives the session that produced it, human review gates that place a named person between a model’s output and the bench, experimental record ownership and data integrity rules that keep the chain unbroken from request to result, peptide identity purity and impurity confirmation backed by orthogonal methods, and activity data read alongside the impurities that could explain it. None of this is new law. The USPTO’s revised inventorship guidance still reserves inventorship to natural persons, and the USPTO’s position that AI systems are tools used by human inventors applies no separate legal standard to them. What moves is the tooling around that stable baseline, which is why the record, not the model, is the asset worth protecting.
Tiết lộ: MOL Changes cung cấp dịch vụ phân tích và tổng hợp peptide tùy chỉnh, so we have a commercial interest in how documentation expectations develop. If you want to see how these pillars translate into practice, review the documentation model or open a technical discussion with the analytical team.